Chapter of Solid state stability

Crystallization tendency

How readily a neat drug crystallizes from its undercooled melt, its glass or a rapidly dried solution, and why.

What this chapter covers

the DSC and spin-coating classifications, critical cooling and heating rates, nucleation and growth windows, crystal growth rates, melt viscosity, thermodynamic driving force and molecular descriptors, polymorph selection from the supercooled liquid, counterion effects in amorphous salts, and solute association in solution before crystallization. The papers and their findings are in sources.md and findings.md.

9 papers2007–201621 findings

01Findings map

From basic to applied. The colour shows the strength of the evidence; click a card to read the finding.

02Which paper supports which finding

Show the table of papers and findings
Kinetics of crystallization in supercooled drug liquidsScreening tests built on themDistinction between forming a glass and keeping itFormulation consequences
paperF1F2F3F4F5F6F7F8F9F10F11F12F13F14F15F16F17F18F19F20F21
2007towler2007solution interactions and crystallization outcome
2008towler2008counterion and amorphous organic salts
2010baird2010DSC classification from undercooled melts
2010vaneerdenbrugh2010aclassification after rapid solvent evaporation
2012baird2012melt viscosity and glass-forming ability
2013baird2013rapid heat–cool calorimetry
2014trasi2014acrystal growth rates of 31 drug liquids
2014trasi2014bnilutamide nucleation far below Tg
2016lu2016vemurafenib polymorphs from the supercooled liquid
other library papers43 papers outside the chapter

Dot size = number of citations of the paper in the finding (long version C). Click a dot to open the finding.

03Connections to other subcategories

Gives to

Solid state stability / Physical stability

The neat-drug class is the starting point for stability work on polymer dispersions: the separated nucleation and growth zones and the class framework were taken up for acetaminophen, with the polymer amount needed tied to glass-forming ability and stability trasi2012a C39, C4; inherent crystallization tendency was named the most important parameter for amorphous tacrolimus, whose neat amorphous form began to crystallize only after 2 weeks at 40 °C/75% RH, with crystallization incomplete after 4 weeks trasi2017 C6, R16; and lumefantrine and artemether were classed with the 2010 screen trasi2020 R5, R6. See F17.

Takes from

Solid state stability / Physical stability

Melt growth rates and their temperature dependence came first from felodipine, bifonazole and nimesulide studies kestur2010b R7; kestur2011 R7, R9, nucleation zones from acetaminophen and flutamide trasi2012a R12; trasi2012b R10, and the growth maximum near 0.94–0.95 Tm was rechecked for other compounds sato2016 E2; tian2017 E3. See F1, F2, F3.

Gives to

Solid state stability / Miscibility and drug-polymer interactions and PEG dispersions

The class of the pure drug was used to choose model drugs and to explain how much polymer is needed vaneerdenbrugh2010b C1, R4 and to read crystallization in PEG dispersions zhu2011 R6, C16; zhu2012 C41. See F17.

Gives to

Dissolution / D. Crystallization from solution and C. Amorphous solubility and LLPS

The solid-state class was tested as a predictor of crystallization from supersaturated solution, with partial success hsieh2012 C42; hsieh2014 C30; ilevbare2013c C24; raina2015b C28, and slow crystallizers were linked to liquid–liquid phase separation ilevbare2013a C27. See F18.

Gives to

Dissolution / B. Supersaturation from ASD dissolution and A. Release mechanism and LoC

Counterion effects on amorphous salts were built on by in situ salt dispersions and by salt–ASD combinations duong2022a C66; hiew2022b R7, C37. See F20.

Gives to

ASD process and Bioperformance

Class III glass formers were chosen for melt-extrusion studies moseson2020 C5, slow crystallizers were judged suitable for solvent granulation trasi2019 C35, and pretomanid and delamanid were placed in class I with reduced Tg 0.67 and 0.64 nguyen2023a C59. See F17.

Left to other subcategories: polymer effects on the stability of amorphous dispersions (Solid state stability / Physical stability; Solid state stability / Miscibility and drug-polymer interactions), crystallization from supersaturated aqueous solution and amorphous solubility (Dissolution / D. Crystallization from solution; Dissolution / C. Amorphous solubility and LLPS), and processing (ASD process). These subcategories have no chapter page yet, so they are named here without links.

How to read this page. Each finding is one conclusion: a headline, a plain statement, one picture and what it means for your drug. Open How it works for the steps and numbers, and Evidence for the data and conditions, at medium (B) or full (C) length. The coloured tag grades the evidence as strong, moderate or weak. Dotted words show a definition. An id such as hate2017 R15 names a paper and an item in its card. Text marked (synthesis: …) joins several papers and is not one paper's claim.

Evidence length

Topic 1 of 4 · 8 findings · F1–F8

Kinetics of crystallization in supercooled drug liquids

F1

Topic 1Kinetics of crystallization in supercooled drug liquids1 of 8

Some drug melts grow crystals ten thousand times faster than others.

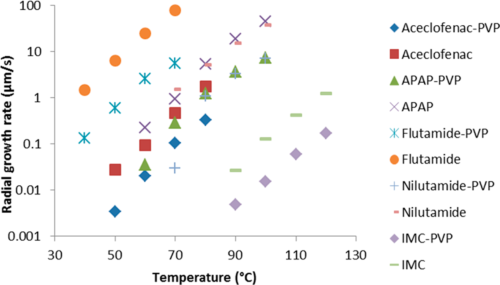
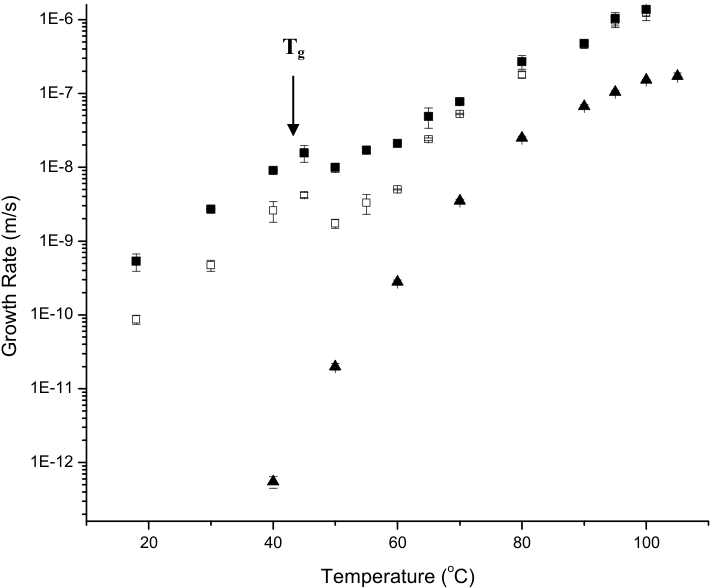
Melt the drug, spread it as a thin film, add a seed crystal and measure under the microscope how far the crystal edge moves each second. Across 31 drugs the fastest edge moved about ten thousand times faster than the slowest trasi2014a R1.a, R1.ae, C23.

moderateOne large data set measured by one method, with no replicates; some maximum values, including the fastest one, disagree with the rates plotted in the same paper trasi2014a X17, X3.

slow · 0.01–1intermediate · 1–10fast · ≥ 10 µm/sclotrimazole 0.02nilutamide ≈40vemurafenib ≈106nimesulide 216≈ 10⁴ × · four orders of magnitude0.010.11101001000highest crystal growth rate of the melt, µm/s (log scale)

Clotrimazole and nimesulide are the extremes of the 31-drug set (the 216 value is doubted by the extractor); nilutamide and vemurafenib come from separate studies. trasi2014a R1.a, R1.ae, C23, X3; lu2016 R12.o; trasi2014b R8.j

For your drug

  1. Measure seeded growth of the crystal form you care about at three or more temperatures between Tg and Tm, and confirm the form by Raman or XRPD.
  2. Place the highest rate in the fast (10 µm/s or more), intermediate (1–10) or slow (0.01–1 µm/s) group, and compare drugs on the same temperature scale, preferably T/Tg.

trasi2014a P6, X2, C6, C7; lu2016 P4

How it works and key numbers
  1. Just below the melting point the molecules move easily, but the driving force for crystallization is small trasi2014a C22.
  2. On cooling the driving force rises while the melt becomes more viscous, so growth is fastest at an intermediate temperature (see F2) trasi2014a C22.
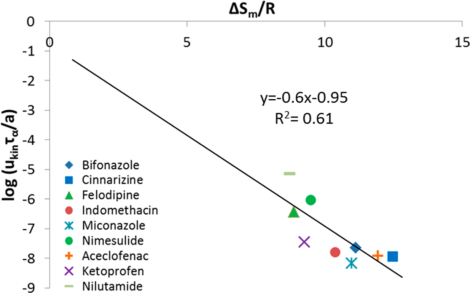
  3. Drugs differ in how viscous their melt is and in their melting entropy; the authors say this "could explain much" of the spread, while other factors "clearly play a role" trasi2014a C38.
  4. Within one drug, cooling the melt toward Tg slows growth by roughly two to six orders of magnitude kestur2010a R5; kestur2011 R7, R9; trasi2014b R8.
To feel the numbers. At 106 µm/s a vemurafenib crystal edge crosses about 1 mm in 10 s (106 × 10 = 1,060 µm); at 0.02 µm/s clotrimazole needs about 14 h for the same distance (1,000 ÷ 0.02 = 50,000 s).
0.02 → 216 µm/sslowest to fastest maximum growth rate among 31 drugstrasi2014a R1.a, R1.ae
2–6 ordersdrop in growth rate when one drug's melt is cooled toward Tgkestur2010a R5; kestur2011 R7, R9; trasi2014b R8
≈ 4 vs ≈ 5 ordersspread between fastest and slowest drug at equal T/Tg vs at equal undercooling; the ranking changes tootrasi2014a R3, C7

Connections to other chapters

Evidence: data, conditions and papers

Finding F1. Crystal growth rates from undercooled drug melts differ by about four orders of magnitude between drugs

In short. Measured by seeded growth in melt films above Tg, the maximum crystal growth velocity (MCV) of 31 drugs ran from 216 µm/s (nimesulide) to 0.02 µm/s (clotrimazole), about four orders of magnitude, and the ranking of drugs changed with the temperature scale used to compare them trasi2014a R1.a, R1.ae, C23, C7. Within one drug, cooling the melt toward Tg lowered the growth rate by roughly two to six orders of magnitude, depending on the drug and the temperature span kestur2010a R5, X1; kestur2010b R7; kestur2011 R7, R9; trasi2014b R8. Vemurafenib and nilutamide are among the fast growers, at about 106 and 40 µm/s lu2016 R12.o; trasi2014b R8.j.

Type. observation in N systems (31 drugs in one study; felodipine, bifonazole, nimesulide, nilutamide, vemurafenib and indomethacin elsewhere) Strength. one large direct data set measured by one method (trasi2014a K1: moderate to strong), but without replicates and with MCV values of unstated origin, several of which disagree with the plotted rates (trasi2014a section 0.8: moderate).

How it was shown

Trasi and co-workers (2014) melted 3–5 mg of each of 31 drugs between coverslips, quenched, seeded the film at the edge and filmed the growth front on a hot stage from about 30 to 90 °C above Tg trasi2014a P1, A1. The highest rate of each drug, its MCV, sorted the drugs into fast (10 µm/s or more), intermediate (1–10) and slow (0.01–1 µm/s) growers trasi2014a P6, X2.

What the data show

MCVs spanned a ratio of about 10⁴, and the rank order of the drugs changed with the temperature scale used trasi2014a R1.a, R1.ae, X14, R3, C7. Within single drugs the temperature dependence is steep kestur2011 C6; trasi2014b R8.

drug setmethod and conditionsresultids
31 neat drugsseeded melt films, 30–90 °C above Tg, no replicatesMCV 216 µm/s (nimesulide) to 0.02 µm/s (clotrimazole); 13, 10 and 8 drugs fast, intermediate, slowtrasi2014a R1, R1.a, R1.ae
31 neat drugssame data on two scalesabout 4 orders at equal T/Tg, about 5 at equal undercooling; order of drugs changestrasi2014a R3, C7
felodipine form IIfilms grown at 70–110 °C, triplicatepeak near 105 °C; about 51-fold fall to 70 °C (text: "3 orders")kestur2010a R5.a, C10, X1
bifonazole; nimesulide18–100 and 18–80 °Cabout 5.5 and 5–6 orders lower at 18 °Ckestur2011 R7, R9
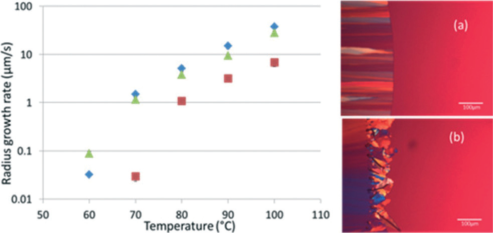
nilutamide stable formseeded films, 30–100 °Cabout 0.00015 µm/s at 30 °C to about 40 µm/s at 100 °Ctrasi2014b R8.a, R8.j
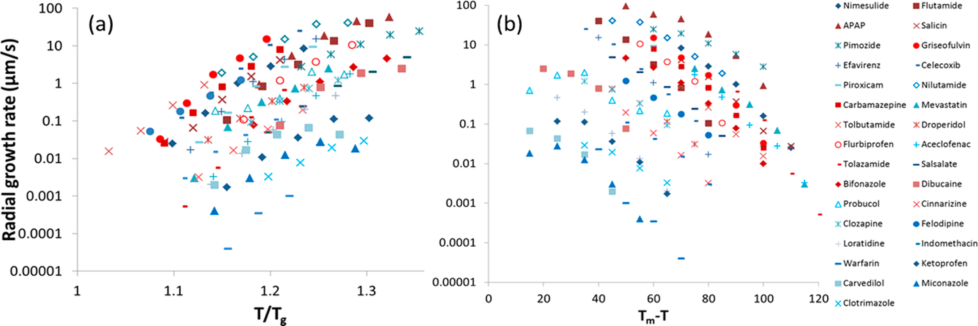
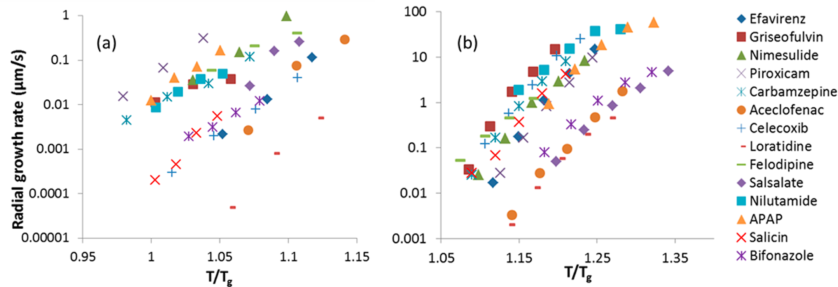
Fig. 1 of trasi2014a: bulk growth rates of 31 drugs against T/Tg and undercooling, coloured by glass-forming class
Fig. 1 · trasi2014abulk growth rates of 31 drugs against T/Tg and undercooling, coloured by glass-forming class

Compare panel (a), against T/Tg, with panel (b), against undercooling, to see the order of drugs change trasi2014a R2, R3.

Why

The authors place growth between two opposing factors: near Tm molecules are mobile but the driving force is small, and on cooling the driving force rises while mobility falls trasi2014a C22. In Jackson's growth law mobility enters as 1/\eta and the bracket is the driving force trasi2014a E1, C31. Differences in melt viscosity and melting entropy "could explain much" of the spread between drugs trasi2014a C38.

Limits and transfer

The 31-drug set has no replicates, the MCV derivation is not stated, and several MCVs lie far from the plotted rates trasi2014a X17, X3. The growing polymorph is assumed from the seed trasi2014a C6. Measure seeded growth of the polymorph of interest at three or more temperatures, confirm its form by Raman or XRPD, and compare drugs at the same T/Tg trasi2014a C7; lu2016 P4.

How it was shown

Kestur and Taylor (2010) grew metastable form II felodipine from melt-quenched films about 30–50 µm thick between cover glasses: the films were nucleated over P2O5 at 40 °C, then the spherulites were grown on a hot stage at 70–110 °C, the slope of diameter against time giving the rate, in triplicate kestur2010a P2, P3, P4, R3. A second 2010 paper used the same two-step protocol on neat felodipine and PVP mixtures kestur2010b P2, P3. Kestur and Taylor (2011) applied it to bifonazole (18–100 °C) and nimesulide (18–80 °C), moving slow samples into ovens or coolers when growth was too slow for one microscopy session kestur2011 P2, P3. Trasi and co-workers (2014) then compared 31 chemically diverse drugs with one method: 3–5 mg of powder was melted between coverslips, quenched, seeded at the edge with the starting crystals so that the polymorph of interest grows, and the front was filmed on a hot stage from about 30 to 90 °C above Tg trasi2014a P1, A1. The highest rate of each drug, its MCV, was used to sort the drugs into fast (10 µm/s or more), intermediate (1–10) and slow (0.01–1 µm/s) growers trasi2014a P6, X2. No replicates or uncertainties are reported, and how the MCVs were obtained is not stated trasi2014a X17, X3. Trasi and Taylor (2014) measured seeded growth of stable and metastable nilutamide in triplicate at 30–100 °C trasi2014b P2, P3, R8. Lu and Taylor (2016) held vemurafenib glass at fixed temperatures from 120 to 270 °C on a hot stage and identified each growing crystal by Raman microscopy lu2016 P2, P4. Tian and co-workers (2017) measured three indomethacin polymorphs from 80 °C to near their melting points, with at least three samples per point tian2017 P3, P4.

What the data show

In the 31-drug set, plotted bulk rates covered about 4 × 10⁻⁵ to 100 µm/s, at T/Tg of about 1.03–1.36 and undercoolings of about 15–120 K trasi2014a R2. MCVs ran from 216 µm/s for nimesulide to 0.02 µm/s for clotrimazole, a ratio of about 10⁴ trasi2014a R1.a, R1.ae, X14. Thirteen drugs had MCVs of 20.6 µm/s or more, ten lay between 1.1 and 8.2 µm/s and eight between 0.02 and 0.86 µm/s trasi2014a R1. Compared at equal T/Tg, the fastest and slowest drugs differed by about four orders of magnitude, with griseofulvin fastest; compared at equal undercooling the spread was about five orders, with acetaminophen and pimozide fastest, so the rank order depends on the normalization trasi2014a R3, C7. Within single drugs the temperature dependence is steep kestur2011 C6; trasi2014b R8. Felodipine form II peaked near 105 °C at about 1.8 × 10⁻⁷ m/s and fell to about 3.5 × 10⁻⁹ m/s at 70 °C, about 51-fold, although the text speaks of 3 orders of magnitude kestur2010a R5.a, C10, X1. The second felodipine study placed the peak at 105 ± 5 °C with a fall of about 2 orders from 110 to 70 °C kestur2010b R7, C9. Bifonazole growth fell about 5.5 orders from 100 to 18 °C and nimesulide 5–6 orders from 80 to 18 °C kestur2011 R7, R9. Stable-form nilutamide grew at about 0.00015 µm/s at 30 °C and about 40 µm/s at 100 °C trasi2014b R8.a, R8.j. Vemurafenib α-needles peaked at about 106 µm/s at 250 °C and β reached about 110 µm/s at 230 °C, while γ, δ and α-brown feathers grew at about 4 µm/s or less lu2016 R12. The α maximum matches the preliminary 106 ± 4 µm/s quoted from the 2014 work lu2016 R25. Indomethacin γ had an MCV of 4.07 × 10⁻⁷ m/s, about an order of magnitude below α at 4.34 × 10⁻⁶ m/s tian2017 R7. (synthesis: where the same drug appears in two papers the values mostly agree: bifonazole at 100 °C, about 4.7 × 10⁻⁶ m/s or 4.7 µm/s kestur2011 R7, against an MCV of 4.0 µm/s trasi2014a R1.s; nimesulide at 80 °C, about 8.5 µm/s kestur2011 R9, against about 8.6 µm/s as the highest plotted bulk rate in 2014 but 216 µm/s as its MCV trasi2014a R5.f, R1.a, which supports the extractor's doubt about that MCV trasi2014a X3.)

papersystemmethod and conditionsresultids
kestur2010afelodipine form II, neatfilms nucleated at 40 °C, grown at 70–110 °C, triplicatepeak near 105 °C, ≈1.8 × 10⁻⁷ m/s; ≈3.5 × 10⁻⁹ m/s at 70 °C (≈51-fold; text "3 orders")R3, R5, C10, X1
kestur2010bfelodipine, neatas abovepeak 105 ± 5 °C; about 2 orders lower from 110 to 70 °CR7, C9
kestur2011bifonazole; nimesulide, neatmelt-quenched films, 18–100 and 18–80 °C, triplicateabout 5.5 orders lower from 100 to 18 °C; 5–6 orders lower from 80 to 18 °CR7, R9
trasi2014a31 neat drugsseeded melt films, hot stage, 30–90 °C above Tg, no replicatesMCV 216 (nimesulide) to 0.02 µm/s (clotrimazole); 13 / 10 / 8 fast, intermediate, slow; about 4 orders on T/Tg, about 5 on undercoolingR1, R2, R3, C7, C23
trasi2014bnilutamide stable formseeded films, 30–100 °C, triplicate≈0.00015 µm/s at 30 °C to ≈40 µm/s at 100 °CR8
lu2016vemurafenib polymorphsisothermal growth, 120–270 °Cα-needles ≈106 µm/s at 250 °C; β ≈110 µm/s at 230 °C; γ, δ, brown feathers ≈4 µm/s or lessR12, R25
tian2017indomethacin γ, αpure melt, 80 °C to near Tm, n \geq 3MCV 4.07 × 10⁻⁷ (γ) against 4.34 × 10⁻⁶ m/s (α)R7

Figure

Fig. 1 of trasi2014a: bulk growth rates of 31 drugs against T/Tg and undercooling, coloured by glass-forming class
Fig. 1 · trasi2014abulk growth rates of 31 drugs against T/Tg and undercooling, coloured by glass-forming class

The rates span about six decades on a log axis; compare panel (a), against T/Tg, with panel (b), against undercooling, to see the order of drugs change trasi2014a R2, R3. Blue symbols (class III glass formers) sit at both the top and the bottom of the plot, the point taken up in F15 trasi2014a R1.

Why

The authors place growth between two opposing factors: near Tm molecules are mobile but the driving force is small, and on cooling the driving force rises while mobility falls, so the rate passes through a maximum (see F2) trasi2014a C22. In Jackson's growth law, U = \frac{2 k_B T}{\pi \lambda^2 \eta}\, f \left[1 - \exp\left(-\frac{\Delta H\,\Delta T}{k_B T\,T_m}\right)\right] \exp\left(-\frac{\Delta S}{k_B}\right), mobility enters as 1/\eta, the bracket is the driving force and the last term the fraction of correctly oriented molecules trasi2014a E1, C31. They assume that the Stokes–Einstein relation holds in their temperature range, so that viscosity describes interfacial diffusion, and note that it fails close to Tg trasi2014a C20, C21. Differences in melt viscosity and melting entropy "could explain much" of the spread between drugs, while other factors "clearly play a role" (see F5 and F6) trasi2014a C38. The steep fall within one drug on cooling is the mobility side of the same balance: for nilutamide the authors read growth over the measured range as under kinetic rather than thermodynamic control ("appears") trasi2014b C6. A cited 1993 study had found a spread of about five decades in MCV among organic compounds trasi2014a R13. The formula behind the maximum is on the Jackson interface-kinetics growth rate page.

Conditions and limits

Neat drugs, seeded or nucleated growth in confined films above Tg, mostly 30–90 °C above Tg trasi2014a P1, A2. The 31-drug set has no replicates or error bars, the MCV derivation is not stated, and several MCVs lie far above (nimesulide, salicin, griseofulvin) or below (felodipine, indomethacin) the plotted rates trasi2014a X17, X3. The polymorph of the growing crystal is assumed from the seed, not checked trasi2014a C6. Felodipine was grown as form II in 2010 but seeded with its starting form in 2014, so the values are not directly comparable kestur2010a R3; trasi2014a P1. Bulk growth below Tg, surface growth (see F21) and growth in dispersions at formulation loadings were not part of the 31-drug comparison trasi2014a C35, C8.

Counterexamples and conflicts

None found in the library for the size of the spread. Surface growth near Tg ranks the drugs differently from bulk growth trasi2014a C8. The 216 µm/s MCV of nimesulide is not supported by the plotted bulk rates of the same paper or by the 2011 nimesulide data (see above) trasi2014a X3; kestur2011 R9.

Transfer check

Measure seeded growth of the polymorph of interest at three or more temperatures between Tg and Tm, confirm the form of the growing crystal by Raman or XRPD, and place the highest rate against the fast (10 µm/s or more), intermediate (1–10) and slow (0.01–1 µm/s) groups trasi2014a P6, X2, C6; lu2016 P4. Compare drugs on the same temperature scale, preferably T/Tg, because the order changes with the normalization trasi2014a C7. Report replicates, since the 2014 values carry no stated uncertainty trasi2014a X17.

History

Kestur measured in 2010 how steeply felodipine growth falls on cooling and in 2011 extended this to bifonazole and nimesulide over 18 °C to 100 °C kestur2010a R5; kestur2010b R7; kestur2011 R7, R9. Trasi in 2014 compared 31 drugs under one method and grouped them by MCV trasi2014a R1, C37. Lu in 2016 confirmed vemurafenib as a fast grower, matching the preliminary 2014 value lu2016 R12, R25. Tian in 2017 showed an order-of-magnitude difference between two polymorphs of one drug tian2017 R7.
F2

Topic 1Kinetics of crystallization in supercooled drug liquids2 of 8

Crystals grow fastest at one temperature a little below the melting point, and more slowly both above and below it.

Plotted against temperature, the growth rate is bell-shaped. Where the peak was measured it lay at about 0.90–0.96 of the melting temperature (in kelvin), close to a cited rule of thumb of 0.94 Tm trasi2014a E8, X5; lu2016 R12.o, E2; tian2017 R7, E3; kestur2010a R5.

moderateModerate: every series that spans the peak is bell-shaped, but the peak position was found in only four or five crystal forms, on 5–10 °C steps, and the 0.94 rule is cited, not derived lu2016 K6; tian2017 P4; trasi2014a E8, X5.

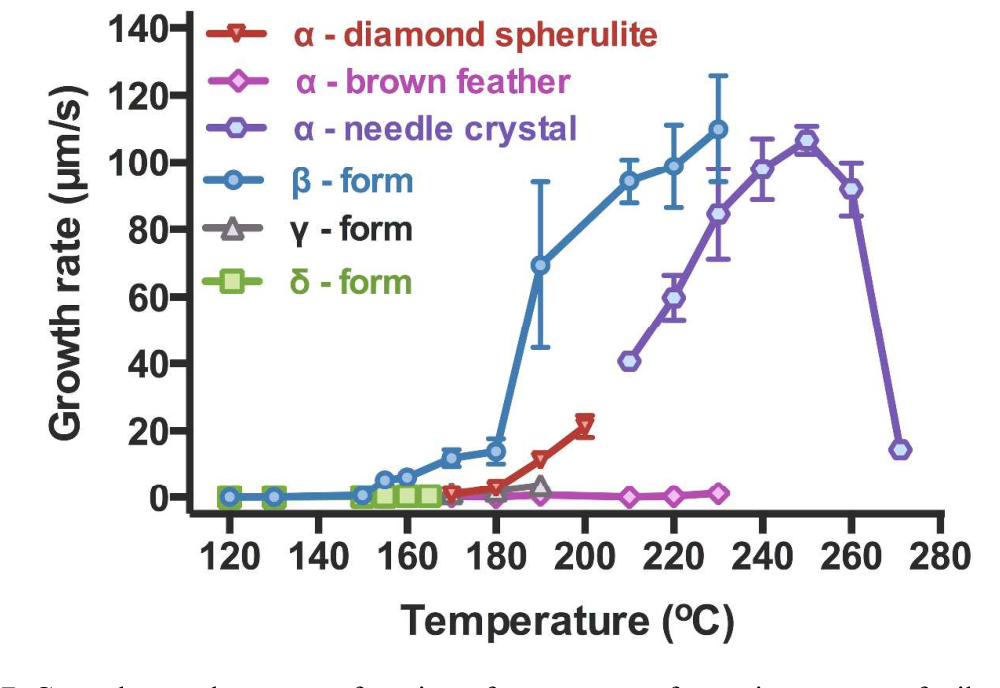
Vemurafenib α-needles: growth rate by temperature, µm/s210 °C41 µm/s220 °C60230 °C85240 °C98250 °C106 · peak260 °C92270 °C14

Growth rises, peaks at 250 °C and then drops steeply toward the melting point at 271 °C. lu2016 R12

vemurafenib αrule 0.940.96indomethacin δ0.940.96indomethacin α0.940.96indomethacin γ0.940.94felodipine form IIagainst the stable-form Tm of 147 °C0.940.900.880.90.920.940.960.981growth peak as a fraction of Tm (kelvin)

Ratios computed in kelvin from the measured peaks; each is uncertain by about ±0.01 because the peaks were located on 5–10 °C steps. lu2016 R12.o, R19.a; tian2017 R7, E3; kestur2010a R5, R3, X7; baird2010 M37; trasi2014a E8

For your drug

  1. Use 0.94 Tm (in kelvin) only to place the window of fastest growth, for example on the cooling path after melt extrusion, where slow cooling leaves the product in the undercooled range.
  2. Measure growth on both sides of the predicted peak before relying on it, and identify the polymorph, since polymorphs of one drug peak at different temperatures.

trasi2014a C34, E8; tian2017 R7; lu2016 R12

How it works and key numbers
  1. Near Tm the molecules move easily, but the driving force for crystallization is small trasi2014a C22; lu2016 C17.
  2. On cooling the driving force rises, roughly in proportion to the undercooling, while the mobility falls trasi2014a C22; lu2016 C17; tian2017 C22.
  3. The product of the two passes through a maximum: above it growth is under thermodynamic control, below it under kinetic (viscosity) control trasi2014a C22; tian2017 C22.
  4. The authors call this "readily explained", but did not measure the mobility of the liquid lu2016 C17.
  5. A check: for indomethacin α, normalizing the growth rate for the driving force removed the maximum; the normalized rate kept rising up to 150 °C tian2017 R11.
Analogy. A skier on a slope that gets steeper but also turns from ice into deep snow: the steeper slope pulls harder, the snow slows the skier, and the best speed is somewhere in the middle.
To feel the numbers. For vemurafenib, Tm = 271 °C = 544.15 K, so 0.94 × 544.15 = 511.5 K = 238.4 °C, about 240 °C. The measured peak was 250 °C, about 10 °C higher.
0.90–0.96 Tmwhere the measured growth peaks lay, in kelvinlu2016 R12.o; tian2017 R7, E3; kestur2010a R5
250 vs 240 °Cvemurafenib α peak as measured vs as given by the 0.94 Tm rulelu2016 R12.o, R24
0.94 TmNaito's rule, cited from the literature and not derived in the librarytrasi2014a E8, X5

Connections to other chapters

1 more connection
Evidence: data, conditions and papers

Finding F2. Crystal growth rate passes through a maximum at roughly 0.90–0.96 of the melting temperature

In short. Growth rate against temperature is bell-shaped, because on cooling the driving force rises while molecular mobility falls trasi2014a C22; lu2016 C17; tian2017 C22. The library uses Naito's rule that the maximum lies near 0.94 Tm in kelvin, cited rather than derived trasi2014a E8, X5; lu2016 E2; sato2016 E2. Where a maximum was measured it lay between about 0.90 and 0.96 Tm: vemurafenib α at 250 °C, indomethacin polymorphs at 110–135 °C and felodipine near 105 °C (synthesis: ratios computed in kelvin in the evidence table) lu2016 R12.o; tian2017 R7, E3; kestur2010a R5.

Type. observation in N systems (vemurafenib, three indomethacin polymorphs, felodipine) against a cited rule Strength. the bell shape is seen in every series that spans the maximum, but its position is located in four or five crystal forms only, on 5–10 °C temperature steps (lu2016 K6: moderate).

How it was shown

Lu and Taylor (2016) measured vemurafenib growth isothermally at 17 temperatures from 120 to about 270 °C and identified each crystal type by Raman lu2016 A1, P2, P4, R19. Tian and co-workers (2017) measured three indomethacin polymorphs from 80 °C to near each melting point, in 5 or 10 °C steps with at least three samples per point tian2017 P4.

What the data show

Vemurafenib α-needles grew at about 41, 60, 85, 98, 106 and 92 µm/s at 210–260 °C, a bell-shaped curve with its maximum at 250 °C, 10 °C above Naito's prediction lu2016 R12, R24, C18. For indomethacin the authors write "approximately 0.95" of the melting temperature tian2017 R7, E3.

formmeasured maximumratio to Tm in kelvinids
vemurafenib α-needles (Tm 271 °C)250 °C, about 106 µm/s0.96 (synthesis: 523.15/544.15)lu2016 R12, R12.o, R19.a
indomethacin δ110 °C0.96 (synthesis)tian2017 R7, E3
indomethacin α135 °C0.96 (synthesis)tian2017 R7, E3
indomethacin γ135 °C0.94 (synthesis)tian2017 R7, E3
felodipine form IInear 105 °C0.90 of the stable-form Tm of 147 °C (synthesis)kestur2010a R5; baird2010 M37
Fig. 7 of lu2016: growth rate against temperature for every vemurafenib form
Fig. 7 · lu2016growth rate against temperature for every vemurafenib form

Follow the purple α-needle series: it rises from 210 °C, peaks at 250 °C and drops steeply toward the melting point lu2016 R12.

Why

Near Tm molecules are mobile but the driving force is small, while on cooling the driving force rises and mobility falls, so the product of the two passes through a maximum trasi2014a C22; lu2016 C17; tian2017 C22. The authors call the bell shape "readily explained" but did not measure the mobility of the liquid lu2016 C17. The 0.94 factor is cited from Naito, not derived trasi2014a E8, X5.

Limits and transfer

Maxima were located on 5–10 °C steps, so a ratio is uncertain by about ±0.01 (synthesis: half of a 5–10 K step divided by 400–550 K) tian2017 P4; lu2016 A1. Use 0.94 Tm in kelvin only to place the window of fastest growth, measure growth on both sides of it, and identify the polymorph trasi2014a E8; tian2017 R7; lu2016 R12.

How it was shown

Locating a maximum needs isothermal growth rates on both sides of it (synthesis: only the papers below measured on both sides or reported the position). Kestur and Taylor (2010) grew felodipine form II spherulites from films nucleated at 40 °C, at 70, 80, 90, 95, 100, 105 and 110 °C, in triplicate kestur2010a P4, R3. A second 2010 felodipine study used the same temperatures kestur2010b P3. Trasi and co-workers (2014) measured 31 drugs from about 30 to 90 °C above Tg, reported one MCV per drug, cited the 0.94 Tm rule and did not tabulate the temperature of each maximum trasi2014a P1, R1, E8, X4. Lu and Taylor (2016) measured vemurafenib growth isothermally at 17 temperatures from 120 to about 270 °C, close to the melting point of 271 °C, identifying each crystal type by Raman lu2016 A1, P2, P4, R19. Tian and co-workers (2017) measured the δ, α and γ polymorphs of indomethacin from 80 °C to close to each melting point in 5 or 10 °C steps, with at least three samples per point tian2017 P4. Sato and co-workers (2016) cited 0.94 Tm to place their measurement temperatures, all of which lay below the maximum sato2016 E2, C14.

What the data show

Vemurafenib α-needles grew at about 41, 60, 85, 98, 106 and 92 µm/s at 210, 220, 230, 240, 250 and 260 °C and at about 14 µm/s near 270 °C, a bell-shaped curve with its maximum at 250 °C lu2016 R12. Naito's rule gives 240 °C for α, 10 °C below the observed maximum lu2016 R24, C18. In kelvin the observed maximum is 0.96 Tm (synthesis: 523.15/544.15) lu2016 R12.o, R19.a. Indomethacin δ, α and γ each gave a bell-shaped curve, with maxima at 110, 135 and 135 °C against onset melting points of 400.15, 425.15 and 433.15 K; the authors write "approximately 0.95" of the melting temperature tian2017 R7, E3. The ratios are 0.96, 0.96 and 0.94 (synthesis: 383.15/400.15, 408.15/425.15, 408.15/433.15) tian2017 E3. Felodipine form II peaked near 105 °C and grew more slowly at 110 °C kestur2010a R5, C10; kestur2010b R7, C9. Against the 147 °C melting point of the stable form, that is 0.90 Tm (synthesis: 378.15/420.15) baird2010 M37, but the crystals that grew were metastable form II, whose own melting point is not given kestur2010a R3, X7. For the 31 drugs no maximum temperatures were reported; the extractor notes that the printed ΔG and ΔSm columns, read with \Delta G \approx \Delta S_m\,\Delta T, imply maxima between 0.90 and 0.99 Tm trasi2014a X4, E3. In nilutamide the stable form's growth was still rising at 100 °C, the highest temperature measured, against a melting point near 153 °C trasi2014b R8.j, M1.

papersystemmethod and conditionsresultids
lu2016vemurafenib α-needles (Tm 271 °C)isothermal growth, 210–≈270 °Cmaximum ≈106 µm/s at 250 °C; 0.94 Tm gives 240 °C; 523.15/544.15 K = 0.96 Tm (synthesis)R12, R19, R24, C18
tian2017indomethacin δ, α, γpure melt, hot stage, n \geq 3maxima at 110, 135, 135 °C against Tm 400.15, 425.15, 433.15 K, "approximately 0.95"; 0.96, 0.96, 0.94 (synthesis)R7, E3
kestur2010afelodipine form II70–110 °C, triplicatemaximum near 105 °C; 0.90 of the stable-form Tm of 147 °C (synthesis)R5, R3, X7
kestur2010bfelodipine70–110 °Cpeak at 105 ± 5 °CR7, C9
trasi2014a31 drugsMCV only0.94 Tm cited, Tcryst,max not reported; printed ΔG and ΔSm imply 0.90–0.99 Tm (extractor)E8, C37, X4, X5
trasi2014bnilutamide (introduction)literaturegrowth maximum "typically" at 0.94 Tm, nucleation maximum below itC3
sato2016six compoundsliterature rulemaximum "typically" near 0.94 Tm; all measurement temperatures below itE2, C14

Figure

Fig. 7 of lu2016: growth rate against temperature for every vemurafenib form
Fig. 7 · lu2016growth rate against temperature for every vemurafenib form

Follow the purple α-needle series: it rises from 210 °C, peaks at 250 °C and drops steeply toward the melting point; the blue β series is faster below 230 °C but was not measured through a maximum lu2016 R12.

Why

The authors' explanation is the same in each paper: near Tm molecules are mobile but the driving force for crystallization is small, while on cooling the driving force rises, roughly in proportion to the undercooling, and mobility falls, so the product of the two passes through a maximum trasi2014a C22; lu2016 C17; tian2017 C22. Above the maximum growth is under thermodynamic control, below it under kinetic (viscosity) control trasi2014a C22; tian2017 C22. The authors call the bell shape "readily explained" but did not measure the mobility of the liquid lu2016 C17. Driving force and mobility are combined in growth laws of the form u = \frac{k}{\eta}\left[1 - \exp\left(-\frac{\Delta G}{RT}\right)\right], used in the felodipine studies kestur2010a E1. Normalizing growth for the driving force removed the maximum for indomethacin α, whose normalized rate kept rising up to 150 °C tian2017 R11. The 0.94 factor itself is an empirical observation cited from Naito, not derived in the library trasi2014a E8, X5. Trasi and co-workers use it to argue that a drug with a small Tm − Tg reaches its maximum closer to Tg and so in a more viscous liquid (see F5) trasi2014a C27. (synthesis: the four measured forms lie within 0.94–0.96 Tm and felodipine form II lower; a fixed ratio of this kind is consistent with growth curves that scale with Tm, but the five forms are too few to test the rule.) The relation is written out on the temperature of maximum crystal growth rate page.

Conditions and limits

Neat melts; maxima located on 5–10 °C temperature steps, so a ratio is uncertain by about ±0.01 (synthesis: half of a 5–10 K step divided by 400–550 K) tian2017 P4; lu2016 A1. The rule is cited, and the 31-drug MCVs come without the temperature at which they occurred trasi2014a X5, X4. Growth with polymers can move the maximum: with 3% polymer the felodipine maximum appeared at 110 °C instead of 105 °C, except with PVAc kestur2010a R5, C11. For vemurafenib, thermal stability was checked only for the 1 min melt-quench step at 285 °C lu2016 P1, R2.

Counterexamples and conflicts

The vemurafenib maximum lay 10 °C above the rule's prediction lu2016 R24, R12.o. The 0.90–0.99 range implied by the 2014 table rests on an unstated ΔG method trasi2014a X4.

Transfer check

Use 0.94 Tm (in kelvin) only to place the window of fastest growth, for example on the cooling path after melt extrusion, where slow cooling leaves the product in the undercooled range trasi2014a C34, E8. Measure growth on both sides of the predicted maximum before relying on it, and identify the polymorph, since polymorphs of one drug peak at different temperatures tian2017 R7; lu2016 R12.

History

The rule enters the library through the 2014 papers trasi2014a E8; trasi2014b C3. The felodipine maximum had been measured in 2010 kestur2010a R5; kestur2010b R7. Vemurafenib in 2016 and indomethacin polymorphs in 2017 gave maxima close to the rule lu2016 R24; tian2017 R7. Sato in 2016 used the rule to place measurement temperatures sato2016 C14.
F3

Topic 1Kinetics of crystallization in supercooled drug liquids3 of 8

Nuclei form at one temperature and crystals grow at another; when these ranges are far apart, the melt forms a glass.

Nucleation (formation of new crystal nuclei) is fastest at deeper cooling, growth closer to the melting point. In the three drugs where both were measured, the two ranges were far apart trasi2012a C35, C22, R6; trasi2012b R10; trasi2014b R5, R8.

moderateMeasured directly for three drugs; for the DSC classes the overlap is only inferred and drawn as schematics baird2010 X5; baird2013 C35.

nucleation range (new nuclei form)growth measured (darker = faster)acetaminophenTg 2450–6070–110flutamide10–4040–70nilutamideTg 32most nuclei at −301.5 → 40 µm/s−40−20020406080100120temperature, °C

Nucleation ranges from cool-and-reheat DSC; growth from hot-stage films; nilutamide nuclei counted after 30 min holds (12.6 at −30 °C against 0.8–2.5 between −10 and 40 °C). trasi2012a C22, R6; trasi2012b R10, R3; trasi2014b R5, R8, R9, M1; baird2010 M24

For your drug

  1. Cool the melt to lower and lower end points, well below Tg, and reheat each time; an exotherm on reheating shows that the end point lay in the nucleation zone. Measure growth separately on seeded films.
  2. Well separated windows predict an easy glass, not a stable one: nuclei made on the way down can grow later (F15, F16).

trasi2012a P6; trasi2014b P1, R5; baird2010 C55; trasi2012a C41

How it works and key numbers
  1. Hot melt: crystals could grow fast here, but few nuclei exist yet baird2013 C36.
  2. Cooler: the melt reaches the window where nuclei form, but it is too cold for them to grow quickly baird2013 C36.
  3. Below Tg: a glass, carrying nuclei that cannot grow in time baird2010 C51.
  4. Reheated: the nuclei pass into the growth window and the glass crystallizes; for acetaminophen the authors think this happens once reheating reaches about 80 °C ("It is thus possible") trasi2012a R6, C38.
Analogy. Seeds that sprout only in the cold and grow only in the warmth: pass quickly through both seasons and you are left with dormant seeds but no plants.
50–60 °Cacetaminophen's narrow nucleation zone; its growth was measured at 70–110 °Ctrasi2012a C22, R6
≈60 °C below Tghow far nilutamide had to be cooled (to −30 °C) before it crystallized on reheatingtrasi2014b R5, M1
3 drugswhere both windows were measured; the class-level picture is inferredbaird2010 X5

Connections to other chapters

Evidence: data, conditions and papers

Finding F3. Nucleation and growth occupy separate temperature windows, and their separation decides whether a melt forms a glass

In short. The library reads crystallization from the melt through two bell-shaped rate curves, nucleation peaking at larger undercooling than growth, and the overlap of the two as what decides glass formation trasi2012a C35; trasi2014a C2; trasi2014b C3. Where both windows were measured they were far apart: acetaminophen nucleated only between 50 and 60 °C while its growth was measured from 70 to 110 °C, flutamide nucleated between 10 and 40 °C, and nilutamide nucleated mainly after cooling to −30 °C while its crystals grew fastest near 100 °C trasi2012a C22, R6; trasi2012b R10; trasi2014b R5, R8. For the crystallization-tendency classes the degree of overlap is inferred from DSC behaviour and drawn as schematics, not measured baird2010 C51, X5; baird2013 C35.

Type. observation in N systems (acetaminophen, flutamide, nilutamide measured) within a framework used across the subcategory Strength. three direct zone measurements and consistent indirect DSC data (trasi2014b K2: moderate to strong), while the class-level reading is inferred (baird2013 K6: weak to moderate; baird2010 K8: weak).

How it was shown

Trasi and Taylor (2012) cooled about 5 mg of acetaminophen melt at 15 °C/min to successively lower end temperatures, held 1 min and reheated at 10 °C/min; the highest end temperature that gave a recrystallization exotherm on reheating marks the upper limit of the nucleation zone trasi2012a P6. Growth was measured separately on a hot stage trasi2012a R6, P7. The same search was applied to flutamide and, with 30 min surface holds, to nilutamide trasi2012b R10; trasi2014b P1, P7.

What the data show

(synthesis: in all three measured drugs the nucleation window lies at lower temperatures than the range of fast growth) trasi2012a C22, R6; trasi2012b R10; trasi2014b R5, R8. The authors call the nilutamide separation "large" trasi2014b C36.

systemnucleation windowgrowth or overlapids
acetaminophen (Tg 24 °C)50–60 °C by DSCfalls nearly two orders from 110 to 70 °Ctrasi2012a C22, R6
flutamidebetween 10 and 40 °Cmeasured at 40–70 °C, falls nearly two orders from 70 to 40 °Ctrasi2012b R10, R3
nilutamide (Tg 32 °C)crystallizes on reheating only after cooling to −30 °C; about 12.6 surface nuclei at −30 °Cabout 1.5 µm/s at 70 °C, about 40 µm/s at 100 °Ctrasi2014b R5, M1, R9, R8
vemurafenib αzone of about 170–270 °C inferred; α never appeared at 165 °C or below; nucleation judged by eyeeach polymorph has its own zonelu2016 C43, R10.a, X8
25 compounds, rapid calorimetryclass II needed reheating rates far above cooling rates; class I-A crystallized even at 1000 °C/minlittle overlap read for class II, strong overlap for class I-Abaird2013 R9, C35, C33
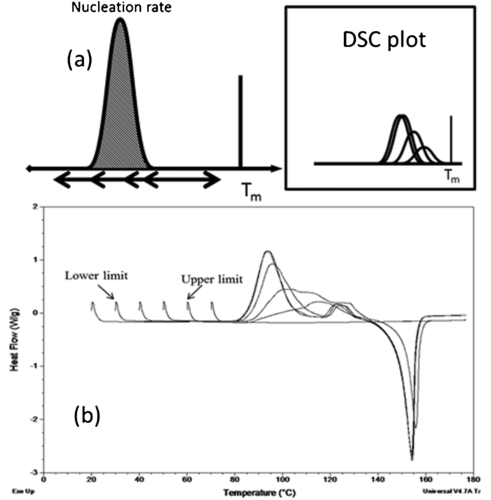
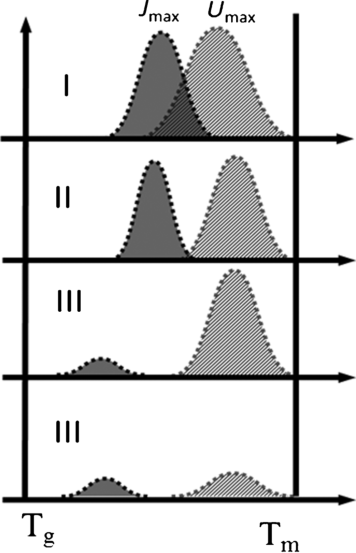
Fig. 1 of trasi2012a: how the overlap of nucleation and growth curves sets the crystallization class
Fig. 1 · trasi2012ahow the overlap of nucleation and growth curves sets the crystallization class

Nucleation rate J (dark) and growth rate U (hatched) are drawn between Tg and Tm, overlapping for class I and separated for class II; it is a schematic, not data trasi2012a C7.

Why

Classical theory gives both rates a maximum between Tg and Tm, nucleation being favoured at larger undercooling trasi2012a C35; trasi2014b C3. Ready glass formation could therefore come from low nucleation, low growth, both, or poor overlap, which a cooling–reheating test alone cannot tell apart trasi2014a C13, C2.

Limits and transfer

Zones were measured for three neat drugs only trasi2012a P6; trasi2014b P1, P7. Nilutamide nuclei counts are counts per sample, not rates trasi2014b X7. Locate the zone by cooling to successively lower end points, extending them well below Tg; separated windows predict glass formation on cooling, not stability on reheating or in storage, because nuclei formed on the way down can grow later trasi2012a P6; trasi2014b R5; baird2010 C55; trasi2012a C41.

How it was shown

Baird and co-workers (2010) proposed the framework for their DSC classes on a time–temperature–transformation diagram adapted from Angell: class I melts cross overlapping nucleation and growth curves and crystallize, class II melts form nuclei that cannot grow before the glass forms, and class III melts pass both curves too fast baird2010 C51, C52. The diagram is a schematic baird2010 X5. Trasi and Taylor (2012) tested the idea on acetaminophen trasi2012a C9. To find the nucleation zone, about 5 mg of melt in a pin-holed pan was cooled at 15 °C/min to successively lower end temperatures, held 1 min and reheated at 10 °C/min: the highest end temperature that gave a recrystallization exotherm on reheating marks the upper limit of the zone, and the end temperature below which the exotherm stops moving marks the lower limit trasi2012a P6. Growth rates were measured separately on a hot stage, and nuclei were counted on a hot stage after one cooling–reheating cycle (mean of 5) trasi2012a R6, P7. The same group applied the DSC zone search to flutamide in 2012 and to nilutamide in 2014, adding for nilutamide 30 min holds of open glass surfaces at −30 to 60 °C followed by 1 min at 65 °C to grow the nuclei for counting trasi2012b R10; trasi2014b P1, P7. Baird and co-workers (2013) inferred the overlap for 25 compounds from the asymmetry between critical cooling and reheating rates in a rapid calorimeter baird2013 C35, R9. Lu and Taylor (2016) inferred a separate nucleation zone for each vemurafenib polymorph from where each form appeared in isothermal crystallization, judging nucleation by eye lu2016 C42, C43, X8.

What the data show

Pure acetaminophen (Tg 24 °C) had a very narrow nucleation zone of 50–60 °C: it had to be cooled to 60 °C or lower to crystallize on reheating trasi2012a C22, R12; baird2010 M24. Its growth rate fell by nearly two orders of magnitude from 110 to 70 °C, so the nuclei formed at 50–60 °C can grow only once reheating reaches about 80 °C ("It is thus possible") trasi2012a R6, C38. The authors conclude that the separated-zone hypothesis "appears to be reasonable" for acetaminophen trasi2012a C39. Flutamide's nucleation zone lay between 10 and 40 °C, and its growth was measured at 40–70 °C, falling nearly two orders of magnitude from 70 to 40 °C trasi2012b R10, R3. Nilutamide crystallized on DSC reheating only after its melt had been cooled to −30 °C, around 60 °C below its Tg of 32 °C trasi2014b R5, M1. Its surface nuclei counts after 30 min holds were about 12.6 at −30 °C, 0.8–2.5 between −10 and 40 °C and about 6 at 50–60 °C, while its stable-form crystals grew at about 1.5 µm/s at 70 °C and about 40 µm/s at 100 °C trasi2014b R9, R8. The authors call this a "large separation" between the temperature ranges favouring nucleation and growth trasi2014b C36. In the rapid calorimeter, class II glasses needed reheating rates far above the cooling rates that formed them, which the authors read as little overlap between the curves; class I-A crystallized even at 1000 °C/min, read as strong overlap baird2013 R9, C35, C33. For vemurafenib the authors place the α nucleation zone at "approximately" 170–270 °C because α never appeared at 165 °C or below ("it appears") lu2016 C43, R10.a. Indomethacin polymorphs likewise nucleated preferentially at different temperatures: δ fastest at 60 °C, α favoured at 80 °C, γ not readily tian2017 R1.

papersystemmethod and conditionsresultids
baird201051 moleculesDSC classes read on a TTT schematicclass I overlapping curves; class II glass with nuclei; class III nuclei-free glass (schematic, not measured)C49, C50, C51, X5
trasi2012aacetaminophenDSC cool–reheat zone search; hot-stage growth and nuclei countsnucleation zone 50–60 °C; growth falls nearly two orders from 110 to 70 °C; separated zones "appears to be reasonable"P6, C22, R6, C38, C39
trasi2012bflutamideDSC cool–reheat; growth at 40–70 °Cnucleation zone between 10 and 40 °CR10, R3
baird20137 class I-B, 10 class II, 8 class I-Arapid heat–cool DSCreheating rates far above cooling rates, read as little overlap; class I-A read as strong overlapR9, C33, C35, C36
trasi2014bnilutamideDSC cool–reheat; 30 min surface holds; seeded growthcrystallization on reheating only after cooling to −30 °C; growth ≈40 µm/s at 100 °C; "large separation"R5, R9, R8, C36
trasi2014a31 drugsgrowth rate against DSC classa cooling–reheating test cannot tell low nucleation from low growth; extended classes IA to IVC2, C13, C17
lu2016vemurafenib polymorphsisothermal crystallization, nucleation judged by eyeeach polymorph has its own nucleation and growth zone; α nucleation zone ≈170–270 °C inferredC42, C43, X8
tian2017indomethacin polymorphsstorage of amorphous filmsδ nucleated fastest at 60 °C; α favoured at 80 °C; γ not readilyR1

Figure

Fig. 1 of trasi2012a: how the overlap of nucleation and growth curves sets the crystallization class
Fig. 1 · trasi2012ahow the overlap of nucleation and growth curves sets the crystallization class

Nucleation rate J (dark) and growth rate U (hatched) are drawn between Tg and Tm: overlapping curves for class I, separated curves for class II, small or widely separated curves for class III; it is a schematic, not data trasi2012a C7.

Fig. 7 of trasi2012a: locating a nucleation zone by DSC
Fig. 7 · trasi2012alocating a nucleation zone by DSC

Panel (b), for acetaminophen with 10% PVPVA, shows the method: no exotherm on reheating after cooling to 70 °C, a recrystallization peak that moves down as the cooling end point is lowered to 30 °C, and no further change below that trasi2012a R11.

Why

Classical crystallization theory gives both rates a maximum between Tg and Tm, nucleation being favoured at larger undercooling and growth closer to the melting point trasi2012a C35; trasi2014b C3. The nucleation maximum arises because the driving force rises and the mobility falls on cooling; near Tg, viscosity "can restrict" diffusion baird2010 C49; trasi2012a C48. To avoid any nucleus, the melt must pass the temperature of shortest nucleation time quickly, R_{Ncrit} \sim \frac{1}{\tau_{Nmin}} with \tau_N = \frac{1}{J_N V}, and nucleation is faster and on a shorter length scale than growth, so the rate that avoids nuclei exceeds the rate that avoids detectable crystals baird2010 E21, E22, C50. In the authors' reading, a melt forms a glass easily when the cooling path crosses the growth window while few nuclei exist and forms nuclei only where growth is slow baird2013 C36. Ready glass formation could therefore come from a low nucleation rate, a low growth rate, both, or poor overlap of the two windows, which a cooling–reheating test alone cannot tell apart trasi2014a C13, C2. For nilutamide the authors attribute the separation to slow nucleation of the stable form, which they link "may be due to" a scarce high-energy conformer in the liquid; conformer populations were not measured (see F7) trasi2014b C51, X15. Each polymorph of a compound may have its own nucleation profile, as for indomethacin γ and α ("it is possible") trasi2014b C38, R32. (synthesis: in all three measured drugs the nucleation window lies at lower temperatures than the range of fast growth, as the class II and class III schemes assume; zones were measured only for drugs that form glasses, because class I melts crystallize before the zones can be separated.) The argument is written out on the critical cooling rate to avoid nucleation and classical nucleation barrier pages.

Conditions and limits

Zones were measured for three neat drugs, from DSC and hot-stage protocols with holds of 1–30 min trasi2012a P6; trasi2012b R10; trasi2014b P1, P7. In the acetaminophen protocols the nucleation zone is passed both on cooling and on reheating, so nuclei from the two steps were not separated trasi2012a X9. Nilutamide nuclei counts are counts per sample from an open surface after 30 min, not rates, and the polymorph of the counted nuclei was not identified trasi2014b X7, X6. Vemurafenib nucleation was judged by eye lu2016 X8. Polymer effects on the zones are a separate topic (acetaminophen's zone broadened with polymers) trasi2012a C22.

Counterexamples and conflicts

None found in the library against separated windows where both were measured. Acetaminophen and flutamide nucleate above Tg, nilutamide mainly far below it, so the separation can lie on either side of Tg (see F4) trasi2014b C37.

Transfer check

Locate the nucleation zone by cooling the melt to successively lower end points and reheating, extending the end points well below Tg, and measure growth separately on seeded films trasi2012a P6; trasi2014b P1, R5. Well separated windows predict glass formation on cooling but not stability on reheating or in storage, because nuclei formed on the way down can grow later (see F15, F16) baird2010 C55; trasi2012a C41.

History

Separated zones were proposed for class II by Baird in 2010 baird2010 C51. Trasi in 2012 tested the idea on acetaminophen and found it reasonable, and measured the flutamide zone trasi2012a C9, C39; trasi2012b R10. Baird in 2013 inferred little overlap for many compounds from the cooling–heating asymmetry baird2013 C35. Trasi in 2014 found an extreme separation in nilutamide and pointed out that the classes cannot separate nucleation from growth trasi2014b C36; trasi2014a C2. Lu in 2016 and Tian in 2017 applied the idea to individual polymorphs lu2016 C42; tian2017 R1.

F4

Topic 1Kinetics of crystallization in supercooled drug liquids4 of 8

A drug glass can nucleate fastest far below Tg, so cold storage does not always protect it.

Nilutamide glass (Tg onset 32 °C) crystallized on reheating only after its melt had been cooled to −30 °C, about 60 °C below Tg. Counting the nuclei on the glass surface after 30 min holds gave a U-shaped curve, highest at −30 °C trasi2014b M1, R5, R9. The authors call this "unusual" trasi2014b C57.

moderateModerate: three tests point the same way, but it is one drug, the counts are not rates (mean of five), and the form of the counted nuclei was not identified trasi2014b K2, X7, X6.

Nilutamide: surface nuclei counted after 30 min−30 °Cabout 60 °C below Tg12.6−20 °C2.3−10 to 40 °C0.8–2.550 °C5.860 °C6.0

Counts per sample, not rates; the −10 to 40 °C bar is drawn at the top of its 0.8–2.5 range. trasi2014b R9, X7

About 50 DSC holds at −30 °C: outcome, % of runsnew metastable form I88%stable form5%no crystals5%form II2%

Holds of 30 min, product identified by Raman; at 60 °C, 94% of the runs did not crystallize. trasi2014b R16

For your drug

  1. Before cold storage, freeze-drying, cryomilling or cold shipment of an amorphous drug, hold glass samples at the lowest temperature they will meet, then reheat or warm them, and identify any crystals by polymorph.
  2. Use holds longer than 30 min where the product will spend days or months in the cold. Nilutamide is a small glass-former that nucleates slowly above Tg; a drug that nucleates readily above Tg, like flutamide, falls outside this evidence.

trasi2014b P8, X16, C37

How it works and key numbers
  1. The usual expectation is that the low mobility of a glass protects it, and storage about 50 °C below Tg has been proposed to prevent crystallization trasi2014b C2, C4.
  2. The authors read the nilutamide counts as two nucleation zones, a strong one far below Tg and a weaker one beginning about 30 °C above Tg, "likely" one per polymorph trasi2014b C41.
  3. They name three possible causes for nucleation of the metastable form in the cold: cracks and stress formed on cooling, control by β-relaxation rather than viscosity, and different molecular conformers (shapes) in the liquid trasi2014b C45, C43, C46.
  4. They favour the conformer idea: the stable crystal contains a higher-energy conformer that should become scarcer in the cold liquid, so the stable form nucleates slowly ("may be due to"). Conformer populations in the liquid were not measured trasi2014b C51, C34, X15.
To feel the numbers. Tg is 32 °C, so −30 °C lies 32 − (−30) = 62 °C below it, which the authors round to about 60 °C. 88% of about 50 holds is about 0.88 × 50 = 44 runs that gave form I.
−30 °Cthe only end temperature that crystallized on DSC reheating, against a Tg of 32 °Ctrasi2014b M1, R5
12.6 vs 0.8–2.5nuclei counted at −30 °C against −10 to 40 °C, 30 min holdstrasi2014b R9
88%of about 50 DSC holds at −30 °C that gave a new metastable form Itrasi2014b R16
Evidence: data, conditions and papers

Finding F4. Nucleation can be fastest far below Tg, so cold storage does not protect every drug glass

In short. Nilutamide glass (Tg onset 32 °C) crystallized on DSC reheating only after its melt had been cooled to −30 °C, about 60 °C below Tg trasi2014b M1, R5. On an open glass surface, 30 min holds gave about 12.6 nuclei at −30 °C against about 0.8–2.5 between −20 and 40 °C, and 88% of about 50 DSC holds at −30 °C produced a new metastable form I trasi2014b R9, R16. The authors call this "unusual", because low-temperature storage is generally considered to stabilize amorphous solids; acetaminophen and flutamide, by contrast, nucleate above their Tg trasi2014b C57, C37.

Type. single-system observation (nilutamide), with cited reports for indomethacin and ibuprofen Strength. DSC, surface counts and DSC holds point the same way (trasi2014b K2: moderate to strong), but counts are not rates, n = 5 and the polymorph of the counted nuclei was not identified (trasi2014b section 0.8: moderate; X7, X6).

How it was shown

Trasi and Taylor (2014) cooled about 5 mg of nilutamide melt at 15 °C/min to a chosen end temperature, held 1 min and reheated at 10 °C/min, lowering the end temperature run by run trasi2014b P1. On an open glass surface, nuclei were counted after 30 min at −30 to 60 °C and 1 min at 65 °C trasi2014b P7, R1. About 50 DSC runs per temperature were held 30 min at −30 or 60 °C and the product was identified by Raman trasi2014b P8.

What the data show

Only the run cooled to −30 °C crystallized on reheating, and the counts and the DSC holds agreed trasi2014b R5, R9, R16.

testconditionsresultids
DSC screencooled to successive end points, reheated at 10 °C/mincrystallization on reheating only after −30 °C, then melting of form I at 141 °Ctrasi2014b P1, R5, R6
surface nuclei counts30 min at −30 to 60 °C, mean of 5about 12.6 at −30 °C; 0.8–2.5 from −20 to 40 °C; about 5.8 and 6.0 at 50 and 60 °C (counts per sample)trasi2014b P7, R9, X7
DSC holds at −30 °Cabout 50 runs, Raman88% form I, 2% form II, 5% stable formtrasi2014b P8, R16
DSC holds at 60 °Cabout 50 runs, Raman94% no crystallizationtrasi2014b P8, R16
acetaminophen; flutamideDSC cool and reheatnucleation zones 50–60 °C and 10–40 °C, both above Tgtrasi2012a C22; trasi2012b R10
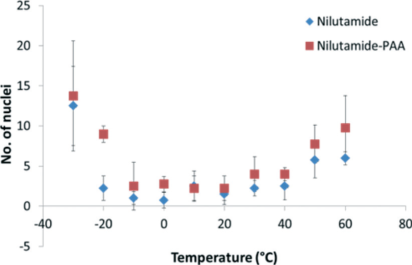
Fig. 5 of trasi2014b: surface nuclei counted after 30 min holds, a U-shaped curve
Fig. 5 · trasi2014bsurface nuclei counted after 30 min holds, a U-shaped curve

Read the blue series (nilutamide): many nuclei at −30 °C, few from −20 to 40 °C and a smaller rise at 50–60 °C; the values are counts per sample, not nucleation rates trasi2014b R9, X7.

Why

The usual expectation is that the low mobility of a glass protects it trasi2014b C2. The authors name three possible causes for favoured nucleation of the metastable form at low temperature: cracks formed on cooling, β-relaxation control, and different conformers or tautomers trasi2014b C45, C43, C46. They favour a conformer route, in which the stable crystal's higher-energy conformer "would be expected" to become scarcer in the cold liquid ("may be due to"); conformer populations were not measured trasi2014b C51, X15.

Limits and transfer

One neat drug; counts per sample for 30 min only, since longer room-temperature storage nucleated both forms trasi2014b X7, X16, R17. No dispersion at a formulation drug loading was tested trasi2014b A4. Before cold storage or cold shipment, hold glass samples at the lowest temperature they will meet, reheat, and identify any crystals by polymorph (synthesis) trasi2014b P8.

How it was shown

Trasi and Taylor (2014) asked why nilutamide, a small drug with a single rotatable bond, forms a glass so readily, and at which temperatures and into which forms its liquid or glass nucleates trasi2014b M1, A1. In a DSC screen, about 5 mg in a pin-holed hermetic pan was heated to 160 °C, cooled at 15 °C/min to a chosen end temperature, held 1 min and reheated at 10 °C/min; the end temperature was lowered run by run until crystallization appeared on reheating trasi2014b P1. For nucleation counts, about 3 mg was melted and cooled between a slide and a coverslip, the coverslip was removed to expose the surface (where nucleation is much faster than in the bulk), the glass was held 30 min at a temperature between −30 and 60 °C, then 1 min at 65 °C to grow the nuclei to visible size, and the nuclei were counted (mean of 5) trasi2014b P7, R1. To identify what nucleates, about 50 DSC runs per temperature were held 30 min at −30 or 60 °C and reheated at 10 °C/min, and the product was identified by Raman trasi2014b P8. Growth rates of the stable form and of the new form I were measured on seeded films at 30–100 °C trasi2014b P2, P3. Comparison data came from the same group's earlier zone measurements on acetaminophen and flutamide trasi2012a C22; trasi2012b R10.

What the data show

In the DSC screen only the run cooled to −30 °C showed an exotherm on reheating (onset about 88 °C, peaks about 97 and 104 °C), followed by melting at 141 °C, the melting point of a polymorph different from the starting material trasi2014b R5, R6. A run cooled only to 0 °C gave a glass that did not crystallize on reheating up to melting trasi2014b R2. The surface counts traced a U shape: about 12.6 nuclei at −30 °C, 2.3 at −20 °C, 0.8–2.5 between −10 and 40 °C, and about 5.8 and 6.0 at 50 and 60 °C; above 60 °C the film flowed trasi2014b R9. The DSC holds agreed: at −30 °C, 88% of runs gave form I, 2% form II, 5% the stable form and 5% no crystallization, while at 60 °C 94% did not crystallize and the rest gave the stable form trasi2014b R16. The 5% stable-form outcomes at −30 °C came from samples that had already nucleated and partly crystallized during cooling, at about 100–115 °C trasi2014b R16. Form I melts at 141 °C with about 66 J/g, against about 153 °C and 98 J/g for the stable form, and is about 3.4 kJ/mol less stable at 25 °C trasi2014b R6, M1, R7. Samples kept a few hours at room temperature nucleated both metastable and stable forms trasi2014b R17. The authors read the counts as two nucleation zones, a strong one far below Tg and a weaker one beginning about 30 °C above Tg, "likely" belonging to form I and the stable form trasi2014b C41. In contrast, acetaminophen (Tg 24 °C) nucleated at 50–60 °C and flutamide (Tg 271 K) between 10 and 40 °C, both above Tg trasi2012a C22; baird2010 M24; trasi2012b R10; trasi2014a M2. The paper cites two indomethacin results: glass stored 55 °C below Tg gave a small melting endotherm on reheating, while the nucleation maxima of its γ and α forms lie at about Tg and about 20 °C higher; a metastable ibuprofen form nucleates when held around 60 °C below Tg trasi2014b R34, R32, R33.
papersystemmethod and conditionsresultids
trasi2014bnilutamide meltDSC cooling at 15 °C/min to successive end points, reheating at 10 °C/mincrystallization on reheating only after cooling to −30 °C, then melting of form I at 141 °CP1, R5, R6
trasi2014bnilutamide glass, open surface30 min at −30 to 60 °C, then 1 min at 65 °C; mean of 5≈12.6 nuclei at −30 °C; ≈0.8–2.5 from −20 to 40 °C; ≈5.8 and ≈6.0 at 50 and 60 °C (counts per sample)P7, R9, X7
trasi2014bnilutamide, DSC holds30 min at −30 or 60 °C, about 50 runs each, Raman−30 °C: 88% form I, 2% form II, 5% stable form; 60 °C: 94% without crystallizationP8, R16
trasi2014bnilutamide, room temperaturestorage of melt-quenched samples for a few hoursboth metastable and stable forms nucleatedR17
trasi2014bindomethacin; ibuprofen (cited)literatureindomethacin glass 55 °C below Tg gave a melting endotherm on reheating; a metastable ibuprofen form nucleates about 60 °C below TgR34, R33
trasi2012aacetaminophen (Tg 24 °C)DSC cool–reheatnucleation zone 50–60 °C, above TgC22
trasi2012bflutamide (Tg 271 K)DSC cool–reheatnucleation zone 10–40 °C, above TgR10

Figure

Fig. 5 of trasi2014b: surface nuclei counted after 30 min holds, a U-shaped curve
Fig. 5 · trasi2014bsurface nuclei counted after 30 min holds, a U-shaped curve

Read the blue series (nilutamide): many nuclei at −30 °C, few from −20 to 40 °C and a smaller rise at 50–60 °C; the values are counts per sample after a 30 min hold and 1 min at 65 °C, not nucleation rates trasi2014b R9, X7.

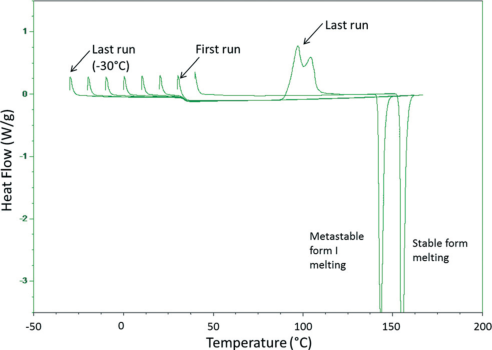
Fig. 3 of trasi2014b: DSC reheating traces after cooling the melt to successively lower temperatures
Fig. 3 · trasi2014bDSC reheating traces after cooling the melt to successively lower temperatures

Only the last run, cooled to −30 °C, shows a crystallization exotherm on reheating, followed by melting of form I trasi2014b R5.

Why

The usual expectation is that the low molecular mobility of a glass protects it, and storage about 50 °C below Tg has been proposed as a good way to prevent crystallization trasi2014b C2, C4. The authors' reading is that nilutamide has two nucleation zones, one per polymorph, as each polymorph may have its own nucleation profile ("it is possible") trasi2014b C38, C41. They name three possible causes for favoured nucleation of the metastable form at low temperature: cracks and mechanical stress formed on cooling (postulated for ibuprofen), control of nucleation far below Tg by β-relaxation rather than viscosity (Oguni), and different conformers or tautomers in the liquid trasi2014b C45, C43, C46. Only the tautomer idea was tested, and a 1H–15N correlation showing the NH intact in form I ruled it out trasi2014b C48, R26. They favour the conformer route: the stable crystal holds a higher-energy conformer (rings at 54.4°) that "would be expected" to become scarcer in the liquid on cooling, so the stable form nucleates slowly and form I, assumed to hold a lower-energy conformer, nucleates at low temperature ("may be due to") trasi2014b C51, C34, C55. Conformer populations in the liquid were not measured at any temperature, and the explanation is one of several untested ones trasi2014b X15. They also reject Ostwald's rule of stages as the source of the stable-form nuclei above Tg, because form I needs about 24 h to convert at 70 °C, far longer than the growth step ("somewhat unlikely") trasi2014b C44, R23. (synthesis: whatever the cause, the nilutamide data show that the minimum of nucleation counts lay between −20 and 40 °C, that is from about 50 °C below to 10 °C above Tg, so the "50 °C below Tg" rule sits at the edge of the deep-cold nucleation zone for this drug trasi2014b R9, M1, C4.) For ASDs, Moseson and Taylor 2023 (p. 6) give the same rule as a rule of thumb for a stability period of years and note reported ASDs with long-term physical stability under much less stringent conditions.

Conditions and limits

One neat drug; 30 min holds; counts per sample without a stated area, from an open surface, compared with sealed DSC pans trasi2014b X7. The low counts between −20 and 40 °C hold for the 30 min time scale only, since longer room-temperature storage nucleated both forms trasi2014b X16, R17. The polymorph of the counted nuclei was not identified trasi2014b X6. Only two polymers at low levels were studied, and the effect of PVP on nucleation could not be measured trasi2014b C22. No amorphous solid dispersion at a formulation drug loading was tested trasi2014b A4.

Counterexamples and conflicts

Acetaminophen and flutamide nucleate above Tg trasi2012a C22; trasi2012b R10. The nucleation maxima of two indomethacin polymorphs lie at or above Tg trasi2014b R32. With 10% molar PAA the nilutamide count at −20 °C was about 9.0 against 2.3, so the polymer effect on nucleation is unresolved rather than absent trasi2014b X5, R9.

Transfer check

Before cold storage, freeze-drying, cryomilling or cold shipment of an amorphous drug, hold glass samples at the lowest temperature they will meet, then reheat or warm them, and identify any crystals by polymorph (synthesis: the nilutamide DSC-hold protocol applied to a process question) trasi2014b P8. Use holds longer than 30 min where the product will spend days or months in the cold, because the 2014 counts cover 30 min only trasi2014b X16. A drug inside the evidence is a small glass-former that nucleates slowly above Tg; a drug that nucleates readily above Tg, like flutamide, falls outside it trasi2014b C37.

History

The rule of storing about 50 °C below Tg is cited in the library trasi2014b C4. Trasi and Taylor in 2014 reported nilutamide as a counterexample, nucleating a previously unreported metastable polymorph far below Tg, and placed it beside earlier indomethacin and ibuprofen reports trasi2014b C10, C13, R33, R34. No later library paper revisited the question.

F5

Topic 1Kinetics of crystallization in supercooled drug liquids5 of 8

Low-viscosity melts crystallize readily and viscous melts form glasses, but viscosity alone cannot rank drugs within a class.

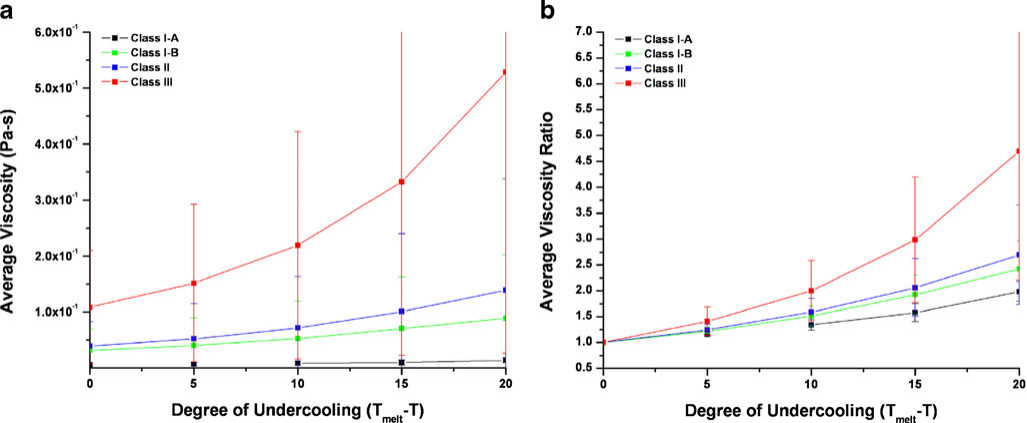
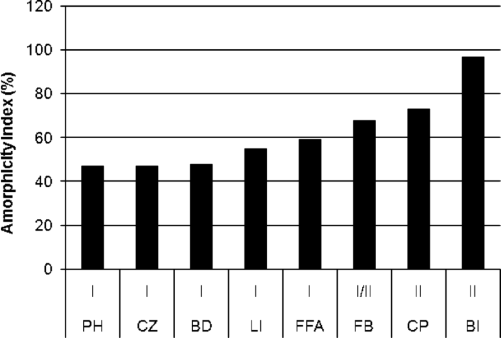
Viscosity is the resistance of the melt to flow. At the melting point, all six melts of the fastest crystallizers (class I-A) had viscosities below 0.01 Pa·s, and all sixteen melts of the slowest (class III) lay above it. Inside a class, viscosity did not rank the drugs baird2012 R1, C22, C11, C16.

moderateModerate: direct viscosity data under one protocol and a clean split of the two extreme classes, but the class means have wide spreads, runs were single, and the mechanism was not measured baird2012 K1, K2, K3, X12, X15.

Class I-Afastest crystallizers0.01 Pa·s2.86–8.62 × 10⁻³Class IIIslowest, stay amorphous0.01 Pa·s13.34 × 10⁻³ to 1.990.0010.010.1110melt viscosity at Tm, Pa·s (log scale)

The two extreme classes do not overlap; the dashed tick marks 0.01 Pa·s. baird2012 R1, C22

Viscosity rise over 20 °C of cooling, class averageClass I-Afastest2.0×Class I-B2.4×Class II2.7×Class IIIslowest4.7×

Class I-B and II did not differ significantly. baird2012 R9, R10

For your drug

  1. Measure the viscosity at Tm and its rise over the first 20–40 °C of undercooling in a parallel-plate rheometer, remelting between steps. A melt below about 0.01 Pa·s at Tm flags rapid crystallization on cooling; one above it whose viscosity rises 35-fold or more over 40 °C resembles the class III compounds.
  2. Confirm by DSC, and compare the driving force before concluding from viscosity alone; Tm − Tg from one DSC run is a cheap proxy at class level only.

baird2012 P1, C46, R3, C25, X9; trasi2014a R9, C26

How it works and key numbers
  1. Classical theory says the nucleation rate falls with the energy needed for viscous flow, and screw-dislocation growth is inversely proportional to viscosity, so a viscous melt should crystallize more slowly baird2012 E1, E3, C4.
  2. The authors therefore expect a faster viscosity rise on cooling to hinder diffusion of molecules to form nuclei; near Tm, class I-A melts are mobile and the driving force is small, so they crystallize "primarily under thermodynamic control" baird2012 C32, C33, C9.
  3. For growth, a drug with a high melt viscosity at Tm and a short range down to Tg is more viscous still at its fastest-growth temperature (near 0.94 Tm), so it grows more slowly trasi2014a C27.
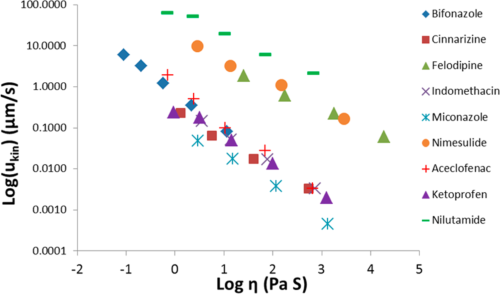
  4. But viscosity is not the whole story: at similar viscosity, growth rates corrected for the driving force still differed by 2–3 orders of magnitude, and melting entropy explains part of the rest (see F6) trasi2014a C30, C32.
  5. The authors call viscosity "indeed one critical attribute", but an exact correlation "is not to be expected in all cases" baird2012 C48, C34.
Undercooling reached before crystallizing, °Ctolazamide6.61 × 10⁻³ Pa·s at Tm≈105celecoxib51.30 × 10⁻³ Pa·s at Tm≈45

The less viscous melt (tolazamide) reached a larger undercooling than the more viscous one (celecoxib), contrary to the simple rule. baird2012 R2, R1, C11, C16

0.01 Pa·sviscosity at Tm: all six class I-A melts lie below it, all sixteen class III melts abovebaird2012 R1, C22
2.0× vs 4.7×viscosity rise over 20 °C of cooling, class I-A against class III averagebaird2012 R9
2–3 ordersgrowth-rate gap between drugs of similar viscosity, after correcting for the driving forcetrasi2014a C30, R10

Connections to other chapters

Evidence: data, conditions and papers

Finding F5. Melt viscosity and the Tm − Tg gap separate the fastest from the slowest crystallizers at class level, but do not rank drugs within a class

In short. All six class I-A melts had viscosities below 0.01 Pa·s at Tm (2.86–8.62 × 10⁻³ Pa·s) and all sixteen class III melts lay above it (13.34 × 10⁻³ to 1.99 Pa·s); on average, viscosity also rose faster on cooling for the slower classes baird2012 R1, R9, C46. Slow growers likewise had viscous melts and a small Tm − Tg, and across 17 drugs viscosity at Tm fell linearly with Tm − Tg (R^2 = 0.7368) trasi2014a R8.k, R8.h, R9. Within classes viscosity did not rank compounds, and at similar viscosity, growth rates corrected for driving force still differed by 2–3 orders of magnitude baird2012 C11, C16; trasi2014a C30.

Type. observation in N systems (34 drugs with viscosity; 31 drugs with growth rates) Strength. direct viscosity data under one protocol with a clean split of the extreme classes (baird2012 K1, K2: moderate; K3: strong), but an association between class means with wide spreads, single runs, and a mechanism that was not measured (baird2012 section 0.8: moderate; X15).

How it was shown

Baird and co-workers (2012) measured 34 of the 51 compounds of the 2010 DSC screen (6 class I-A, 5 I-B, 7 II, 16 III) in a rheometer with 25 mm parallel plates, shearing at 30 s⁻¹ from Tm downward in 5 °C steps and remelting between steps to remove nuclei baird2012 A1, P1. Trasi and co-workers (2014) took the viscosities of 17 of their 31 drugs from this study and compared them with growth rates trasi2014a P7, P6.

What the data show

Melt viscosities spanned almost three orders of magnitude, and the extreme classes separated cleanly at 0.01 Pa·s baird2012 R1, C22. The difference between classes I-B and II was "small and not statistically significant" baird2012 R10, C24.

levelcomparisonresultids
classes at Tmηmelt of 34 drugsclass I-A 2.86–8.62 × 10⁻³ Pa·s; class III 13.34 × 10⁻³ to 1.99 Pa·s; class averages about 0.005, 0.032, 0.040, 0.11 Pa·s (I-A, I-B, II, III)baird2012 R1, R8
classes on coolingviscosity ratio over 20 °C of undercoolingabout 2.0, 2.4, 2.7 and 4.7; I-B and II not significantly differentbaird2012 R9, R10
within classesundercooling reached before crystallizationtolazamide about 105 °C against celecoxib about 45 °Cbaird2012 R4, R2, C11, C16
growth, 17 drugsTm − Tg against viscosity at Tmslow group about +0.95 SD in viscosity and −0.99 SD in Tm − Tg; \eta = -0.0028\,(T_m - T_g) + 0.3989, R² 0.7368trasi2014a R8, R9, E6
growth at similar viscositycorrected for driving force, 9 drugsnear \log \eta \approx 1, nilutamide about 20 µm/s against miconazole about 0.019 µm/strasi2014a R10, C30
Fig. 3 of baird2012: class-average viscosity and viscosity ratio against undercooling
Fig. 3 · baird2012class-average viscosity and viscosity ratio against undercooling

Panel (a) keeps the order class III above II above I-B above I-A at every step from 0 to 20 °C; the bars are the spread between compounds of a class, not measurement error baird2012 R8, R9.

Why

In classical nucleation theory the nucleation rate falls with the activation energy for viscous flow, and screw-dislocation growth is inversely proportional to viscosity baird2012 E1, E3, C4. The authors therefore expect a faster viscosity rise to lower the nucleation rate "all other factors being equal" baird2012 C32. They conclude that viscosity is "indeed one critical attribute", while an exact correlation "is not to be expected in all cases" baird2012 C48, C34.

Limits and transfer

Single runs at one shear rate on neat melts; nucleation and growth were not measured baird2012 X12, X15. Antipyrin, 4-biphenylcarboxaldehyde and tolazamide lie below 0.01 Pa·s yet form glasses baird2012 X8. Measure viscosity at Tm and its rise over 20–40 °C of undercooling, and confirm by DSC baird2012 P1, X9.

How it was shown

Baird and co-workers (2012) asked whether the viscosity of a drug's undercooled melt, at Tm and in its rise on cooling, follows its crystallization class baird2012 A1. They measured 34 of the 51 compounds of the 2010 DSC screen (6 class I-A, 5 I-B, 7 II, 16 III) in an Anton Paar MCR 301 rheometer with 25 mm parallel plates and a 0.4 mm gap baird2012 A1, P1. About 0.5 g of powder was melted at 10 °C above Tm, cooled rapidly to Tm and sheared at 30 s⁻¹; before each lower temperature, in 5 °C steps, the sample was remelted at Tm + 10 °C for 5 min to remove nuclei, and a run ended when viscosity rose with time at constant temperature, the sign of crystallization baird2012 P1. Replicates and equilibration times are not stated baird2012 X12. Viscosity at Tm (ηmelt), the ratio \eta/\eta_{melt} and class averages over 0–20 °C of undercooling were compared baird2012 P3, P4, E7. Trasi and co-workers (2014) took the viscosities of 17 of their 31 drugs from the 2012 study, grouped the drugs by maximum growth velocity, and expressed each group's mean property as a deviation from the overall mean in standard deviations trasi2014a P7, P6, E5. For nine drugs with viscosity–temperature data they divided the growth rate by the driving-force factor, U_{kin} = \frac{U}{1 - e^{-\Delta G/RT}} with \Delta G \approx \Delta S_m\,\Delta T, and plotted it against viscosity trasi2014a E2, E3, P8.

What the data show

Melt viscosities spanned almost three orders of magnitude baird2012 R1. Class I-A ran from 2.86 × 10⁻³ (benzamide) to 8.62 × 10⁻³ Pa·s (haloperidol), class III from 13.34 × 10⁻³ (ibuprofen) to 1.99 Pa·s (ritonavir) baird2012 R1. Taking 10⁻² Pa·s as a benchmark, no class I-A compound lay above it and every class III compound did baird2012 C22. Classes I-B and II were mostly above it, with antipyrin, 4-biphenylcarboxaldehyde (I-B) and tolazamide (II) below baird2012 R11, C22. By the tables, 9 of 34 compounds are below 0.01 Pa·s and 3 of these form glasses on quenching baird2012 X3, R11. Class-average viscosity at Tm was about 0.005, 0.032, 0.040 and 0.11 Pa·s for classes I-A, I-B, II and III, and over 20 °C of undercooling it rose about 2.0-, 2.4-, 2.7- and 4.7-fold baird2012 R8, R9. The difference between classes I-B and II was "small and not statistically significant" baird2012 R10, C24. At 40 °C of undercooling, five class III compounds had risen 35- to 87-fold (felodipine 71, ketoprofen 61, loratadine 87, probucol 82, procaine 35), against about 29 for dibucaine, the highest in class II, and about 10 for chlorpropamide (I-B) baird2012 R3, C25. Within classes the ranking failed: benzamide, the least viscous class I-A melt, undercooled further than 4-biphenylmethanol before crystallizing, and tolazamide (6.61 × 10⁻³ Pa·s) reached about 105 °C of undercooling while the more viscous celecoxib (51.30 × 10⁻³ Pa·s) crystallized at about 45 °C baird2012 R4, R2, R1, C11, C16. In the growth data, the slow group lay about +0.95 standard deviations above average in viscosity and −0.99 in Tm − Tg, the fast group −0.73 and +0.49 trasi2014a R8. Across 17 drugs, viscosity at Tm followed \eta = -0.0028\,(T_m - T_g) + 0.3989 Pa·s with R² 0.7368 trasi2014a R9, E6. Bifonazole, aceclofenac and tolazamide grow at intermediate rates although their melt viscosities (0.0101, 0.0143 and 0.0066 Pa·s) and Tm − Tg (134, 143 and 154) sit with the fast growers trasi2014a R9, M16, M17, M19. Corrected for driving force, near \log \eta \approx 1 nilutamide grew at about 20 µm/s and miconazole at about 0.019 µm/s, two sets about 2–3 orders apart trasi2014a R10, C30.

papersystemmethod and conditionsresultids
baird201234 drugs (6 I-A, 5 I-B, 7 II, 16 III)steady shear at 30 s⁻¹ from Tm downward, remelting between steps, single runsηmelt 2.86–8.62 × 10⁻³ Pa·s (I-A) and 13.34 × 10⁻³ to 1.99 Pa·s (III); 9 compounds below 0.01 Pa·s, 3 of them glass formersP1, R1, R11, X3, X8
baird2012class averages0–20 °C undercoolingviscosity ratio at 20 °C ≈2.0 (I-A), ≈2.4 (I-B), ≈2.7 (II), ≈4.7 (III); I-B and II not significantly differentR9, R10
baird2012within classesrheometrybenzamide undercooled further than 4-biphenylmethanol; tolazamide reached ≈105 °C undercooling while celecoxib crystallized at ≈45 °CR4, R2, C11, C16
baird2012probucol, chlorpropamidesimilar ηmelt (8.03 and 9.33 × 10⁻² Pa·s)ratio at 40 °C ≈82 against ≈10; classes III and I-BC33, R3
trasi2014a31 drugs; viscosities of 17 cited from baird2012MCV groups, group deviationsslow group ≈+0.95 SD in viscosity and ≈−0.99 SD in Tm − Tg; \eta(T_m) = -0.0028\,(T_m - T_g) + 0.3989, R² 0.7368R8, R9, E6, P7, X9
trasi2014a9 drugsgrowth corrected for driving force against viscositynear \log \eta \approx 1, nilutamide ≈20 µm/s against miconazole ≈0.019 µm/s; two sets about 2–3 orders apartR10, C30, X15

Figure

Fig. 3 of baird2012: class-average viscosity and viscosity ratio against undercooling
Fig. 3 · baird2012class-average viscosity and viscosity ratio against undercooling

Panel (a) keeps the order class III above II above I-B above I-A at every step from 0 to 20 °C, and panel (b) shows class III rising fastest; the bars are the spread between compounds of a class, not measurement error baird2012 R8, R9.

Fig. 8 of trasi2014a: growth rate corrected for driving force against viscosity, nine drugs
Fig. 8 · trasi2014agrowth rate corrected for driving force against viscosity, nine drugs

Each drug falls steeply as its melt becomes more viscous, but nilutamide, nimesulide and felodipine lie 2–3 decades above the other six at the same viscosity trasi2014a R10.

Why

In classical nucleation theory the nucleation rate falls exponentially with the sum of a thermodynamic barrier and the activation free energy for viscous flow, I_{SS} = I_0 \exp\left[-\frac{W^* + \Delta G_\eta}{k_B T}\right], and screw-dislocation growth is inversely proportional to viscosity baird2012 E1, E3, C4. The authors therefore expect a faster viscosity rise to lower the nucleation rate "all other factors being equal", and read the probucol–chlorpropamide pair as a faster rise that "thus hindered" diffusion of molecules to form nuclei baird2012 C32, C33. For class I-A they read crystallization near Tm as "primarily under thermodynamic control", mobility being high and the driving force small baird2012 C9. Angell's critical melt viscosity of 0.01–1 Pa·s for a good glass former is cited, and the authors find it a "good predictor" for these compounds baird2012 C29, C31. For growth, Trasi and co-workers argue that if all melts had the same viscosity at Tg, a drug with high viscosity at Tm and a short range to Tg would be more viscous at its MCV temperature, near 0.94 Tm, and so grow more slowly ("This makes sense") trasi2014a C27. The three intermediate growers with fluid melts are "most likely" more fragile, and tolazamide "could be" slowed by its structure; neither was measured trasi2014a C29, X18. Nucleation and growth rates, through which viscosity is argued to act, were not measured in the 2012 study baird2012 X15. Competing explanations are in the library: the probucol–chlorpropamide comparison is not controlled for driving force, since probucol's Hoffman ΔGv at 298 K is −13.2 against −26.4 J/cm³ baird2012 X9. Melting entropy accounts for part of the spread left at equal viscosity (see F6) trasi2014a C32. The authors conclude that viscosity is "indeed one critical attribute", while an exact correlation "is not to be expected in all cases" baird2012 C48, C34. The fitted relation is on the linear fit of melt viscosity on Tm − Tg page and the temperature dependence of viscosity on the Vogel-Tammann-Fulcher equation page.

Conditions and limits

Neat melts at one shear rate, with single runs of unstated reproducibility; classes and Tg values come from unsheared DSC, and Tg is estimated for all class I-A compounds baird2012 X12, X13. Shear raised class I-A crystallization temperatures above the DSC values baird2012 R5, C8. The class III average at Tm leaves out ritonavir without saying so baird2012 X6. The 2014 viscosity bars rest on 17 drugs, only 5 of the 13 fast growers trasi2014a X9. Diffusion to nuclei or crystal faces was not measured baird2012 X15.

Counterexamples and conflicts

Antipyrin, 4-biphenylcarboxaldehyde and tolazamide lie below 0.01 Pa·s yet form glasses, and bifonazole sits at the threshold baird2012 X8. Bifonazole, aceclofenac and tolazamide grow at intermediate rates although their melt viscosities are as low as those of fast growers trasi2014a R9, C29. At similar viscosity, growth corrected for driving force differs by 2–3 orders of magnitude, and the upper set includes felodipine, a slow grower by MCV trasi2014a C30, X15.

Transfer check

Measure the viscosity at Tm and its rise over the first 20–40 °C of undercooling in a parallel-plate rheometer, remelting between steps baird2012 P1. A melt below about 0.01 Pa·s at Tm flags rapid crystallization on cooling, and one above it whose viscosity rises 35-fold or more over 40 °C resembles the class III compounds compared in 2012 (synthesis: the named class III values were 35–87, against at most about 29 in class II) baird2012 C46, R3, C25. Confirm by DSC, and compare the driving force before concluding from viscosity alone baird2012 X9. Tm − Tg from one DSC run is a cheap proxy at class level only trasi2014a R9, C26.

History

Angell's critical melt viscosity of 0.01–1 Pa·s is cited baird2012 C29. Baird in 2012 first measured melt viscosities against crystallization class in the library baird2012 R1. Trasi in 2014 linked the same viscosities and Tm − Tg to growth rates and showed the residual spread trasi2014a R9, R10.

F6

Topic 1Kinetics of crystallization in supercooled drug liquids6 of 8

The driving force for crystallization explains only part of the differences between drugs, and sometimes it points the wrong way.

In two 2010 screens, the rapid crystallizers had on average a larger heat of fusion and a larger driving force for crystallization (the Hoffman ΔGv, the free-energy difference between the supercooled liquid and the crystal) baird2010 R22, C61; vaneerdenbrugh2010a R20, C38. In matched pairs the driving force pointed the right way for nifedipine against felodipine, but the wrong way for indomethacin against felodipine and for the vemurafenib polymorphs marsac2006a R4, C15; alonzo2010 C42; lu2016 C52.

weakWeak to moderate: the class averages and a nine-drug fit are consistent, but the causal readings rest on estimated quantities, not measured ones vaneerdenbrugh2010a K6; trasi2014a K4; baird2010 C60, C61.

class I: fast crystallizersclass III: glass formersheat of fusion−0.63+0.45entropy of fusion−0.59+0.41ΔGv (more negative = larger driving force)−0.40+0.57−0.8−0.6−0.4−0.200.20.40.60.8class mean, in standard deviations from the overall mean

51 molecules classified from undercooled melts; fast crystallizers lie on the side of larger heat of fusion, larger entropy of fusion and larger driving force. baird2010 R22

Larger driving force for crystallization
nifedipineindomethacinvemurafenib α
Actually crystallized faster or first
nifedipinefelodipinevemurafenib β

marsac2006a R4, C15; alonzo2010 C42; lu2016 C52

For your drug

  1. Compute the Hoffman ΔGv at your storage or process temperature from the DSC Tm and ΔHfus (per volume, with the crystal density), and use it to compare drugs of otherwise similar viscosity and Tg, not as a stand-alone predictor.
  2. Where storage lies far below Tm, check the estimate by MDSC; for a polymorphic drug, do not expect the form with the larger driving force to appear first.

baird2010 E25, P8; marsac2006a C15, C14; almeidaesousa2015 C8; lu2016 C52

How it works and key numbers
  1. In classical nucleation theory the barrier to form a nucleus is W^* = 16\pi\gamma^3/(3\Delta G_v^2). For a similar interfacial energy γ, a more negative ΔGv gives a lower barrier and faster nucleation baird2010 E24, C61.
  2. The authors add that a lower heat and entropy of fusion imply a smaller driving force at lower temperature, and a higher melting point means more energy to break the lattice baird2010 C60, C62, E23.
  3. For growth, the Jackson factor \exp(-\Delta S/k_B) stands for the fraction of molecules that arrive in the right orientation to join the crystal. A larger melting entropy then means slower growth at equal mobility and driving force trasi2014a C31, C32.
  4. Beware: the screens find rapid crystallizers high in entropy of fusion (per volume), while the growth analysis finds fast growers low in it (per mole). Different bases and read-outs, so melting entropy is not a dependable single predictor baird2010 R22; trasi2014a R8.f.
  5. For nifedipine against felodipine the enthalpy dominates, and the orientation view would predict the opposite order marsac2006a C34, C35.
Analogy. A ball on a slope: a steeper slope helps it roll, but rough ground and an awkward shape matter too.
To feel the numbers. Nifedipine's free-energy difference at 25 °C is 9.6 \div 6.2 = 1.5 times felodipine's, and its nucleation rate is 0.39 \div 0.17 = 2.3 times higher.
9.6 vs 6.2 kJ/molfree-energy difference between amorphous and crystalline drug at 25 °C, nifedipine against felodipine (MDSC)marsac2006a R4.k, R4.l
R² 0.61fit of normalized growth rate against melting entropy, nine drugstrasi2014a R11
−11.0 vs −6.3 kJ/molvemurafenib α against β driving force near 115 °C; β dominated anywaylu2016 R22, C51, C52

Connections to other chapters

Evidence: data, conditions and papers

Finding F6. Thermodynamic driving force and melting entropy explain only part of the differences between drugs

In short. In both 2010 screens, rapid crystallizers had on average larger heats and entropies of fusion per volume and a more negative Hoffman ΔGv, which the authors read through classical nucleation theory as a lower nucleation barrier baird2010 R22, C61; vaneerdenbrugh2010a R20, C38, C39. The ΔGv trend is not monotonic across the three classes vaneerdenbrugh2010a X10. For growth, melting entropy correlated negatively with growth normalized for viscosity and driving force (R² 0.61, nine drugs), read as an orientation penalty, with other factors unaccounted for trasi2014a R11, C31, C32. In matched comparisons the driving force pointed the right way for nifedipine against felodipine but the wrong way for indomethacin against felodipine and for the vemurafenib polymorphs marsac2006a R4, C15; alonzo2010 C42; lu2016 C52.

Type. observation in N systems (51 and 40 compounds; 9 drugs for entropy; matched pairs) Strength. to moderate — consistent class averages and a nine-drug fit, but the causal readings rest on estimated quantities (vaneerdenbrugh2010a K6: weak to moderate; trasi2014a K4: moderate).

How it was shown

The Hoffman free-energy difference \Delta G_v = -\frac{\Delta H_{fus}\,(T_m - T)\,T}{T_m^2} was computed from DSC melting data baird2010 E25; vaneerdenbrugh2010a E2. Baird and co-workers (2010) and Van Eerdenbrugh and co-workers (2010) included ΔHfus, ΔSfus and ΔGv among seven descriptors in principal component analyses of 51 and 40 compounds, reporting class means in standard deviations baird2010 P8, R22; vaneerdenbrugh2010a P11, R20. Trasi and co-workers (2014) plotted corrected growth rates of nine drugs against ΔSm/R trasi2014a R11.

What the data show

In the melt classes, class I averaged about −0.40 SD in ΔGv and class III about +0.57 baird2010 R22. For growth, \log(U_{kin}\,\tau_\alpha/a) fell with ΔSm/R (R² 0.61) trasi2014a R11, E7. Nifedipine's larger driving force than felodipine's is enthalpic marsac2006a C15.

papersystemresultids
baird201051 molecules, melt classesclass I ΔHfus ≈+0.45, ΔSfus ≈+0.41, ΔGv ≈−0.40 SD; class III ≈−0.63, ≈−0.59, ≈+0.57baird2010 R22, C60, C61
vaneerdenbrugh2010a40 drugs, solvent classesΔGv ≈−0.29 (I), ≈+0.29 (II), ≈+0.13 (III)vaneerdenbrugh2010a R20, C38, C39, X10
trasi2014a9 drugs, normalized growth against ΔSm/Rslope −0.6, R² 0.61; fast-growth group ΔSm ≈−0.38 SDtrasi2014a R11, E7, R8.f
marsac2006anifedipine, felodipineΔG ≈−30 against ≈−18 J/g at 0 °C; nifedipine nucleates faster, the difference is enthalpicmarsac2006a R4, R11, C15
lu2016vemurafenib α, βα has the larger Hoffman driving force at every temperature, yet β dominates at 120–190 °Clu2016 R22, C51, C52
Fig. 9 of trasi2014a: normalized growth rate against melting entropy, nine drugs
Fig. 9 · trasi2014anormalized growth rate against melting entropy, nine drugs

A falling trend with considerable scatter (R² 0.61); drugs at similar ΔSm/R still differ by more than a decade trasi2014a R11, C32.

Why

In classical nucleation theory a more negative ΔGv "would result", for a similar interfacial energy, in a lower barrier and faster nucleation; the interfacial energy was not measured baird2010 E24, C61; vaneerdenbrugh2010a C39. For growth, a larger melting entropy means a smaller fraction of molecules with the right orientation, so slower growth at equal mobility and driving force; entropy "certainly plays a role" trasi2014a C31, C32.

Limits and transfer

ΔGv is a Hoffman estimate, not a measurement, except for the MDSC values of nifedipine and felodipine marsac2006a R11; baird2010 C60, C61. The entropy fit rests on nine drugs trasi2014a X11. Use ΔGv to compare drugs of otherwise similar viscosity and Tg, not alone baird2010 E25, P8; marsac2006a C15.

How it was shown

The driving force for crystallization of an amorphous drug is the free energy difference between the supercooled liquid and the crystal, estimated in the library almost always with the Hoffman equation, \Delta G_v = -\frac{\Delta H_{fus}\,(T_m - T)\,T}{T_m^2}, from DSC melting data baird2010 E25; vaneerdenbrugh2010a E2. Marsac and co-workers (2006) compared amorphous nifedipine and felodipine, two dihydropyridines of nearly equal Tg (45.5 and 46.4 °C midpoint), measuring nucleation in films at 22 °C and 0% RH and the free energy difference by modulated DSC from 0 to 100 °C marsac2006a R4, R1, R11. Baird and co-workers (2010) and Van Eerdenbrugh and co-workers (2010) included ΔHfus, ΔSfus and ΔGv at 293.15 K, all per unit volume, among seven descriptors in principal component analyses of 51 and 40 compounds, and reported each class's mean as a deviation from the overall mean in standard deviations baird2010 P8, R22; vaneerdenbrugh2010a P11, R20. Trasi and co-workers (2014) tested melting entropy against growth: for nine drugs they divided the growth rate by the driving-force factor (\Delta G \approx \Delta S_m\,\Delta T) and by \tau_\alpha/a, with \tau_\alpha = \frac{C\eta}{G}, to remove viscosity, and plotted the result against ΔSm/R trasi2014a E2, E3, E4, R11. Lu and Taylor (2016) computed Hoffman free energies for vemurafenib α and β from their melting data lu2016 P7, E1.

What the data show

Nifedipine nucleated faster than felodipine at 22 °C (0.39 against 0.17 × 10¹⁰ nuclei per m³ per day) and crystallized on heating at 87 °C, while felodipine did not crystallize on heating marsac2006a R4.a, R1, R5. Nifedipine's free energy difference was larger at every temperature (about −30 against −18 J/g at 0 °C; 9.6 against 6.2 kJ/mol at 25 °C by MDSC), and MDSC agreed well with the Hoffman estimate (8.8 and 6.2 kJ/mol) marsac2006a R11, R4.k, R4.l, C14. Nifedipine also had the larger heat (39.9 against 30.8 kJ/mol) and entropy of fusion (89.6 against 74.4 J/mol/K), so its larger driving force is enthalpic and in spite of a larger entropy loss marsac2006a R4.g, R4.h, C15. In the 2010 melt classes, class I averaged about +0.45 SD in ΔHfus, +0.41 in ΔSfus and −0.40 in ΔGv, class III about −0.63, −0.59 and +0.57 baird2010 R22. In the 2010 solvent classes, the ΔGv contributions were about −0.29, +0.29 and +0.13 for classes I, II and III, so class II, not class III, had the smallest driving force vaneerdenbrugh2010a R20, X10. For growth, \log(U_{kin}\,\tau_\alpha/a) fell with ΔSm/R as y = -0.6\,x - 0.95 (R² 0.61), from about −5.1 for nilutamide at ΔSm/R ≈ 8.7 to about −8.2 for miconazole at ≈10.9 trasi2014a R11, E7. The fast-growth group had a lower melting entropy than average (about −0.38 SD) and the slow group a higher one (about +0.30) trasi2014a R8.f. In 2012, probucol and chlorpropamide, compared for their viscosity rise, differed twofold in ΔGv at 298 K (−13.2 against −26.4 J/cm³) baird2012 X9. For vemurafenib, α had the larger Hoffman driving force at every temperature (about −11.0 against −6.3 kJ/mol near 115 °C), yet metastable β dominated crystallization between 120 and 190 °C lu2016 R22, C51, C52.

papersystemmethod and conditionsresultids
marsac2006anifedipine, felodipineMDSC and Hoffman, 0–100 °C; films at 22 °C, 0% RHΔG ≈−30 against ≈−18 J/g at 0 °C; nifedipine nucleates faster and crystallizes on heating at 87 °C, felodipine does not; difference enthalpicR4, R11, R1, R5, C15
baird201051 molecules, melt classesclass deviations, energies per volume, ΔGv at 293.15 Kclass I ΔHfus ≈+0.45, ΔSfus ≈+0.41, ΔGv ≈−0.40 SD; class III ≈−0.63, ≈−0.59, ≈+0.57R22, C60, C61
vaneerdenbrugh2010a40 drugs, solvent classessame descriptorsΔGv ≈−0.29 (I), ≈+0.29 (II), ≈+0.13 (III); ΔHfus ≈+0.41, −0.30, −0.26R20, C38, C39, X10
trasi2014a9 drugs\log(U_{kin}\,\tau_\alpha/a) against ΔSm/Rslope −0.6, R² 0.61; fast-growth group ΔSm ≈−0.38 SDR11, E7, R8, C31, C32
baird2012probucol, chlorpropamideHoffman ΔGv at 298 K−13.2 against −26.4 J/cm³, so the viscosity comparison between them is not controlled for driving forceX9
alonzo2010indomethacin, felodipineamorphous solids during dissolutionindomethacin has the larger driving force but crystallizes much more slowlyC42
lu2016vemurafenib α, βHoffman from melting dataα larger driving force at every temperature; β dominates at 120–190 °CR22, C51, C52

Figure

Fig. 7 of marsac2006a: free energy difference between amorphous and crystalline nifedipine and felodipine against temperature
Fig. 7 · marsac2006afree energy difference between amorphous and crystalline nifedipine and felodipine against temperature

The nifedipine points (circles) lie below the felodipine points (triangles) at every temperature, and the Hoffman lines run close to the MDSC points, which is why the Hoffman estimate is used as the driving force elsewhere in the library marsac2006a R11, C14.

Fig. 9 of trasi2014a: normalized growth rate against melting entropy, nine drugs
Fig. 9 · trasi2014anormalized growth rate against melting entropy, nine drugs

A falling trend with considerable scatter (R² 0.61); drugs at similar ΔSm/R still differ by more than a decade trasi2014a R11, C32.

Why

The authors' nucleation argument uses the classical barrier W^* = \frac{16\pi\gamma^3}{3\,\Delta G_v^2}: a more negative ΔGv "would result", for a similar interfacial energy γ, in a lower barrier and faster homogeneous nucleation baird2010 E24, C61. Van Eerdenbrugh and co-workers call the class correlation "consistent with theory" and infer ("suggests") that differences in γ between compounds are small compared with those in ΔGv; γ was not measured and is "difficult to either measure or estimate" vaneerdenbrugh2010a C39. Baird and co-workers add that lower ΔHfus and ΔSfus imply, for equal ΔCp, a smaller driving force at lower temperature, and that a higher melting point means more energy to disrupt the lattice baird2010 C60, C62, E23. For growth, the Jackson factor \exp(-\Delta S/k_B) represents the fraction of molecules with the right orientation to join the crystal, so a larger melting entropy means slower growth at equal mobility and driving force; entropy "certainly plays a role", but other factors, "could include" interactions in the melt and at the interface trasi2014a C31, C32. Marsac and co-workers cite the same orientation view of the entropy of crystallization, but note that it would predict felodipine to crystallize more easily, which it did not; for that pair the enthalpy dominates marsac2006a C34, C35. (synthesis: the two screens express ΔHfus and ΔSfus per volume and find the rapid crystallizers high in both, while the 2014 growth analysis uses ΔSm per mole and finds the fast growers low in it; the opposite signs come from different bases and read-outs, and show that melting entropy is not a dependable single predictor baird2010 R22; trasi2014a R8.f.) The Hoffman estimate itself becomes less accurate at large undercooling, because it assumes a constant heat-capacity difference and a temperature close to Tm, and the screens evaluate it at 293.15 K for drugs melting up to 272 °C almeidaesousa2015 C8; mosqueragiraldo2015 C24; baird2010 P8, M21. The formulas are on the Hoffman equation, classical nucleation barrier, entropy of fusion and linear fit of normalized growth rate on melting entropy pages.

Conditions and limits

ΔGv is a Hoffman estimate, not a measurement, except for the MDSC values of nifedipine and felodipine; interfacial energy, ΔCp(T) and nucleation rates were not measured in the screens marsac2006a R11; baird2010 C60, C61; vaneerdenbrugh2010a C39. Per-compound descriptor values are not printed for the solvent classes, and the class bars are read from plots vaneerdenbrugh2010a R20. The entropy fit rests on nine drugs, with constants whose units are not given trasi2014a X11. Felodipine and nifedipine are not matched in mobility: felodipine's melt viscosity at Tm is about four times nifedipine's (synthesis: 154 against 36.2 × 10⁻³ Pa·s) baird2012 R1.u, R1.q.

Counterexamples and conflicts

Indomethacin has the larger driving force than felodipine yet crystallizes much more slowly during dissolution alonzo2010 C42. Vemurafenib β dominates although α has the larger driving force (see F7) lu2016 C52. ΔGv is least favourable for the intermediate, not the slow, solvent class vaneerdenbrugh2010a X10. A molecular dispersion in PVP lowers nifedipine's driving force less than the difference between nifedipine and felodipine, so the drug itself sets the larger part of the driving force marsac2006b C41.

Transfer check

Compute the Hoffman ΔGv at the storage or process temperature from DSC Tm and ΔHfus (per volume with the crystal density), and use it to compare drugs of otherwise similar viscosity and Tg, not as a stand-alone predictor baird2010 E25, P8; marsac2006a C15. Where the storage temperature lies far below Tm, check the estimate by MDSC as in the nifedipine–felodipine study marsac2006a C14; almeidaesousa2015 C8. For polymorphic drugs, do not expect the form with the larger driving force to appear first lu2016 C52.

History

Marsac in 2006 tied the faster crystallization of nifedipine to its larger, enthalpic driving force marsac2006a C15. Baird and Van Eerdenbrugh in 2010 found the same direction across many compounds baird2010 C40; vaneerdenbrugh2010a C38. Alonzo in 2010 recorded a pair where the driving force pointed the wrong way during dissolution alonzo2010 C42. Trasi in 2014 added melting entropy as an orientation factor for growth trasi2014a C31. Lu in 2016 showed a polymorphic case where the form with the smaller driving force dominates crystallization lu2016 C52.

F7

Topic 1Kinetics of crystallization in supercooled drug liquids7 of 8

The crystal form that appears depends on nucleation and growth rates, not stability, so a less stable form often crystallizes first.

The form that crystallizes first is the one that nucleates and grows fastest at that temperature lu2016 C42, C52; trasi2014b C12. Vemurafenib gave two or three of its four polymorphs together at 120–240 °C, with the metastable β form dominant at 150–190 °C lu2016 R10, C59. Nilutamide nucleated a new metastable form I after cooling to −30 °C trasi2014b R16.

moderateModerate: the forms were identified by several techniques and conversions were seen directly, but the nucleation side of the explanation is qualitative, judged by eye trasi2014b K3, K4; lu2016 K2, K5, X8.

metastable β dominatesstable α predominatesvemurafenib βmetastabledominant 150–190vemurafenib αstableneedles appear 200predominates above 210only α above 250120150180210240270isothermal crystallization temperature, °C

α has the larger driving force at every temperature, yet β dominates between 150 and 190 °C. lu2016 R10, R11, C59, R22, C52

Nilutamide seeded growth, µm/s (log scale)100 °C · stable form40100 °C · form I1160 °C · stable form0.03460 °C · form I0.1430 °C · stable form0.0001530 °C · form I0.005

The stable form is faster at 70–100 °C, the metastable form I at 60 °C and below; the curves cross between 60 and 70 °C. trasi2014b R8, C12

For your drug

  1. Identify crystals formed during processing or stability stress by Raman or XRPD before assuming the stable form; a metastable product can convert later, slowly in the solid state (nilutamide form I about 24 h at 70 °C).
  2. Map growth rates of the known polymorphs at two or more temperatures in the range the product will meet, because the faster form can change with temperature; a minor form can be invisible by PXRD and visible by Raman microscopy.

lu2016 C14, C47; trasi2014b R23, R8

How it works and key numbers
  1. Each form has its own temperature windows for nucleation and growth; the overlap and size of these at the crystallization temperature set which forms appear lu2016 C42.
  2. In vemurafenib, β is limited by nucleation above 150 °C but grows fast into large regions below 200 °C; γ nucleates rapidly but grows slowly; δ nucleates densely near Tg but grows extremely slowly and is engulfed by β. Above 200 °C β nucleation becomes unfavourable and α predominates owing to "more favorable nucleation" lu2016 C46, C47, C48.
  3. Since β dominates although α has the larger driving force, "these observations indicate" that cold crystallization is controlled by factors other than the thermodynamic driving force lu2016 C52, C53.
  4. For nilutamide the authors propose ("may be due to") that the stable crystal's higher-energy conformer becomes scarce in the cold liquid, so form I nucleates and grows faster at low temperature. Conformer populations in the liquid were not measured trasi2014b C51, C34, X15.
  5. Not always: in felodipine the stable form I grew 1–2 orders of magnitude faster than metastable form II over the whole range studied, so there is no crossover kestur2013a C4.
Analogy. Two runners, one quick off the blocks and one faster later: who leads depends on where in the race you look, here at which temperature the material spends its time.
150–190 °Crange where metastable vemurafenib β dominated, although α has the larger driving forcelu2016 R10, C59, R22, C52
60–70 °Cwhere the growth rates of nilutamide's stable form and form I crosstrasi2014b R8, C12
8 moleculesclass I molecules of the 51-compound screen that gave a different polymorph after liquid-N2 quenchingbaird2010 R3, C16

Connections to other chapters

Evidence: data, conditions and papers

Finding F7. The polymorph that crystallizes from a supercooled drug liquid follows nucleation and growth kinetics, often a metastable form at large undercooling

In short. Which form crystallized depended on temperature through each form's nucleation and growth rates rather than on which form is most stable lu2016 C42, C52; trasi2014b C12. Vemurafenib gave two or three of four polymorphs together at 120–240 °C, with metastable β dominant at 150–190 °C although α has the larger driving force at every temperature lu2016 R10, C59, R22, C52. Nilutamide nucleated a new metastable form I after cooling to −30 °C, and the growth rates of form I and the stable form crossed between 60 and 70 °C trasi2014b R16, R8, C12. Eight class I molecules of the 51-compound screen gave a different polymorph after liquid-N2 quenching baird2010 R3, C16.

Type. observation in N systems (vemurafenib, nilutamide, indomethacin, felodipine; eight molecules of the 51-compound screen) Strength. forms identified by several techniques and conversions observed directly (trasi2014b K3, K4: strong; lu2016 K2: moderate to strong), while the nucleation side of the explanation is qualitative (lu2016 K5: weak to moderate; X8).

How it was shown

Lu and Taylor (2016) crystallized vemurafenib glass isothermally at 120–270 °C, identified every crystal type in place by Raman microscopy and measured each form's growth rate lu2016 P2, P4, R12. Trasi and Taylor (2014) held nilutamide melts for 30 min at −30 or 60 °C in about 50 DSC runs each and measured seeded growth at 30–100 °C trasi2014b P8, P3, R8.

What the data show

β grew fastest up to 230 °C, from about 12 µm/s at 170 °C to about 110 µm/s at 230 °C, faster than α below that temperature lu2016 R12, C21. Felodipine shows the other pattern: stable form I grew 1–2 orders of magnitude faster than metastable form II kestur2013a C4.

systemconditionsresultids
vemurafenibisothermal crystallization, 120–270 °C; Raman, PXRDβ and δ at 120 °C; β, γ and α at 170–190 °C; only α above 250 °Clu2016 R10, R11, C59
vemurafenib α, βDSC melting data; Hoffman curvesα 271 °C and 114 J/g, β 248 °C and 68 J/g; monotropic; virtual transition 319 °Clu2016 R19, R22, C50, C51
nilutamide30 min DSC holds; seeded growth at 30–100 °C88% form I after −30 °C holds; stable form 3–4 times faster at 70–100 °C, form I faster at 60 °C and belowtrasi2014b R16, R8, C12
felodipine forms I and IIbulk growth, triplicatestable form I faster than form II by 1–2 orderskestur2013a R8, C4
indomethacinstorage of amorphous filmsδ nucleated fastest at 60 °C, α favoured at 80 °C, γ poorlytian2017 R1
Fig. 5 of lu2016: where each vemurafenib form grows and where it converts
Fig. 5 · lu2016where each vemurafenib form grows and where it converts

Read the bars against the Tg–Tm axis: δ grows only near Tg, γ in a narrow middle band, β over most of the range and α from about 170 °C to Tm lu2016 R9.

Why

Lu and Taylor explain the vemurafenib pattern by different nucleation and growth zones for each polymorph, whose overlap and magnitudes set which forms appear ("This can be explained") lu2016 C42. Since β dominates although α has the larger driving force, "these observations indicate" that cold crystallization is controlled by other factors lu2016 C52. For nilutamide the authors propose, with "may be due to", a scarce high-energy conformer in the cold liquid; conformer populations were not measured trasi2014b C51, X15.

Limits and transfer

The drugs were neat, in thin films, and γ and δ rest on morphology and Raman only lu2016 P1, X6; trasi2014b P2. Close to Tm the stable form prevails lu2016 R10; trasi2014b R16. Identify crystals by Raman or XRPD before assuming the stable form, and map growth of known polymorphs at two or more temperatures lu2016 C14; trasi2014b R8, R23.

How it was shown

Baird and co-workers (2010) identified the forms obtained after quenching class I melts in liquid N2 by Raman spectra taken in liquid N2 and, for the DSC runs, by the melting profile of the recrystallized material baird2010 P5, R3, R11. Kestur and Taylor (2013) measured the bulk growth rates of felodipine forms I and II over a range of temperatures, in triplicate kestur2013a R8. Trasi and Taylor (2014) held nilutamide melts for 30 min at −30 or 60 °C in about 50 DSC runs each and identified the product by Raman; they characterized the forms by XRPD, FTIR, Raman and 13C solid-state NMR and measured seeded growth of the stable form and form I at 30–100 °C in triplicate trasi2014b P8, P3, R15, R20, R25, R8. Conversion of form I to the stable form was followed by Raman at 70–90 °C trasi2014b P5, R23. Lu and Taylor (2016) crystallized vemurafenib glass isothermally at fixed temperatures from 120 to 270 °C on a hot stage, identified every crystal type in place by Raman microscopy with a 2.1 µm spot, ran PXRD on isothermal samples, and measured the growth rate of each form lu2016 P2, P4, P5, R12. They related α and β thermodynamically by the heat-of-fusion rule and the Hoffman equation lu2016 P7, E3, E1. Tian and co-workers (2017) stored amorphous indomethacin films at 60 and 80 °C to obtain δ and α, and seeded γ tian2017 R1, P3.

What the data show

Vemurafenib produced four polymorphs in six morphologies lu2016 R7, C41. At 120 °C only β and δ nucleated; at 170–190 °C β, γ and α diamond spherulites crystallized together; α needles appeared at 200 °C and took over above 210 °C; above 250 °C only α grew lu2016 R10, R11. β grew fastest up to 230 °C, from about 12 µm/s at 170 °C to about 110 µm/s at 230 °C, faster than α below that temperature lu2016 R12, C21. α melts at 271 °C with 114 J/g and β at 248 °C with 68 J/g, so by the heat-of-fusion rule the pair is monotropic, with α the stable form; the Hoffman curves give α the larger driving force at every temperature and a virtual transition at 319 °C lu2016 R19, C50, R22, C51. Metastable forms then converted toward α: γ to β above 200 °C, δ to β near 220 °C, and β to α slowly in the solid state, only where it touched α, and fast once β melted (DSC endotherm near 252 °C at 100 °C/min) lu2016 R15, R17, R13, C14, R21. Nilutamide held 30 min at −30 °C gave form I in 88% of runs, form II in 2% and the stable form in 5%, while holds at 60 °C gave the stable form when they crystallized at all trasi2014b R16. The stable form grew 3–4 times faster than form I at 70–100 °C (about 40 against 11 µm/s at 100 °C), while form I was faster at 60 °C and below (about 0.14 against 0.034 µm/s at 60 °C, about 0.005 against 0.00015 µm/s at 30 °C) trasi2014b R8, C12. Form I is monotropic to the stable form and about 3.4 kJ/mol less stable at 25 °C trasi2014b C11, R7. Felodipine shows the other pattern: stable form I grew 1–2 orders of magnitude faster than metastable form II over the whole range studied kestur2013a C4. Of the class I molecules of the 2010 screen, eight (4-biphenylmethanol, anthranilic acid, benzamide, caffeine, carbamazepine, chlorpropamide, haloperidol, tolbutamide) gave a different polymorph by Raman after quenching, and the authors call recrystallization to a metastable form from the undercooled melt "not unexpected" baird2010 R3, C16. Amorphous indomethacin nucleated δ fastest at 60 °C, favoured α at 80 °C and nucleated γ poorly tian2017 R1.

papersystemmethod and conditionsresultids
lu2016vemurafenibisothermal crystallization of the supercooled liquid, 120–270 °C; Raman, PXRDβ and δ at 120 °C; β, γ and α at 170–190 °C; α takes over above 200 °C, only α above 250 °CR10, R11, C59
lu2016vemurafenib α, βDSC melting data; Hoffman curvesα 271 °C and 114 J/g, β 248 °C and 68 J/g; monotropic; virtual transition 319 °CR19, R20, R22, C50, C51
lu2016vemurafenib growthhot stageβ ≈12 µm/s at 170 °C to ≈110 µm/s at 230 °C, faster than α below 230 °CR12, C21
trasi2014bnilutamide30-min DSC holds; seeded growth at 30–100 °C88% form I after −30 °C holds; stable form 3–4 times faster at 70–100 °C, form I faster at 60 °C and below; monotropic, form I ≈3.4 kJ/mol less stable at 25 °CR16, R8, R7, C11
kestur2013afelodipine forms I and IIbulk growth, triplicatestable form I faster than form II by 1–2 orders over the range studiedR8, C4
baird201023 class I moleculesDSC; Raman in liquid N2 after quench8 gave a different polymorph by Raman; metastable form "not unexpected"R3, R11, C16
tian2017indomethacinstorage of amorphous filmsδ nucleated fastest at 60 °C; α favoured at 80 °C; γ did not nucleate readilyR1
trasi2014bindomethacin γ and α (cited)literaturenucleation maxima at about Tg (γ) and about 20 °C higher (α)R32

Figure

Fig. 5 of lu2016: where each vemurafenib form grows and where it converts
Fig. 5 · lu2016where each vemurafenib form grows and where it converts

Read the bars against the Tg–Tm axis: δ grows only near Tg, γ in a narrow middle band, β over most of the range and α from about 170 °C to Tm; the arrows show the conversions δ to β, γ to β and β to α lu2016 R9.

Fig. 4 of trasi2014b: growth rates of stable nilutamide and form I against temperature
Fig. 4 · trasi2014bgrowth rates of stable nilutamide and form I against temperature

The two curves cross between 60 and 70 °C: the stable form (blue) is faster at 70 °C and above, form I (red) at 60 °C and below trasi2014b R8, X3.

Why

Lu and Taylor explain the vemurafenib pattern by different nucleation and growth zones for each polymorph, whose overlap and magnitudes at the crystallization temperature set which forms appear ("This can be explained") lu2016 C42. In their reading, β is nucleation-limited above 150 °C but grows fast into large regions below 200 °C; γ "nucleates rapidly but grows slowly"; δ nucleates densely near Tg but grows extremely slowly and is engulfed by β; above 200 °C β nucleation becomes unfavourable and α predominates owing to "more favorable nucleation" lu2016 C46, C47, C48. Since β dominates although α has the larger driving force, "these observations indicate" that cold crystallization is controlled by factors other than the thermodynamic driving force, consistent with metastable forms being more probable at larger undercooling lu2016 C52, C53. Nucleation was judged from microscope impressions rather than measured, and γ and δ were never isolated pure lu2016 X8, X6. They cite Ostwald's rule of stages and Yu's point that a fast-nucleating, slow-growing form can be overtaken by a slow-nucleating, fast-growing one lu2016 C37. For nilutamide the authors propose, with "may be due to", that the stable crystal's higher-energy conformer (rings at 54.4°) becomes scarce in the cold liquid, so form I nucleates and grows faster at low temperature trasi2014b C51, C34. Conformer populations were not measured trasi2014b X15. They find Ostwald's rule "somewhat unlikely" as the source of the stable form above Tg, because form I converts too slowly at 70 °C (about 24 h) to be an unobserved intermediate step trasi2014b C44, R23. (synthesis: the four drugs give two patterns: in felodipine the stable form grows faster at all temperatures studied, while in nilutamide and vemurafenib a metastable form grows faster in the colder part of the range; which form appears then depends on where in the undercooled range the material spends its time kestur2013a C4; trasi2014b R8; lu2016 R12.) Monotropy is assigned with the heat-of-fusion rule on the Burger-Ramberger heat-of-fusion rule page, and the driving forces with the Hoffman equation.

Conditions and limits

Neat drugs in thin films lu2016 P1; trasi2014b P2. Vemurafenib growth rates come without a stated number of crystals, and temperature ranges for the same form differ by about 10 °C between figures and text lu2016 X9, X3. γ and δ rest on morphology and Raman only, without PXRD patterns, melting points or structures lu2016 X6. The structure of nilutamide form I was not solved trasi2014b R29. The 2010 polymorph calls disagree between two tables for five compounds baird2010 X2.

Counterexamples and conflicts

Close to Tm the stable form prevails: vemurafenib above 250 °C and nilutamide held at 60 °C lu2016 R10; trasi2014b R16. In felodipine the stable form grows faster at all temperatures studied, so no crossover occurs there kestur2013a C4. The 2016 conclusion orders γ and δ although the paper states their relationship could not be established lu2016 X7.

Transfer check

Identify crystals formed during processing or stability stress by Raman or XRPD before assuming the stable form; a metastable product can convert later, slowly in the solid state (vemurafenib β to α only at α contacts; nilutamide form I about 24 h at 70 °C) lu2016 C14; trasi2014b R23. Map growth rates of the known polymorphs at two or more temperatures in the range the product will meet, because the faster form can change with temperature trasi2014b R8. A minor form can be invisible by PXRD and visible by Raman microscopy, as γ vemurafenib was lu2016 C47.

History

Baird in 2010 noted metastable forms from undercooled melts baird2010 C16. Kestur in 2013 found the stable felodipine form faster at all temperatures kestur2013a C4. Trasi and Taylor in 2014 found a new nilutamide polymorph at low temperature and a growth crossover, reported as unusual trasi2014b C10, C12. Lu and Taylor in 2016 mapped four vemurafenib polymorphs against temperature lu2016 C41. Tian in 2017 saw temperature-dependent nucleation of indomethacin polymorphs tian2017 R1.

F8

Topic 1Kinetics of crystallization in supercooled drug liquids8 of 8

The hydrogen-bond motifs of the dissolved molecules did not indicate which crystal form would appear first.

Infrared and Raman spectra of indomethacin and anthranilic acid solutions showed carboxylic acid dimers, even below saturation. In two of three informative cases the hydrogen-bond motif did not match the polymorph that crystallized first towler2007 R13, R17, R9, R18, R19. The authors conclude that solution motifs are "not necessarily indicative" towler2007 C33, C35.

moderateModerate: the key spectral contrasts are visible in the figures, but it rests on two acids and bulk spectra, and the concentrations, solvent subtraction and crystallized forms are poorly documented towler2007 X5, X6, X7, A1.

solid form III33813495solid form IIthe form that crystallized33733474anthranilic acid in toluene33833497336033903420345034803510NH2 stretch band position, cm⁻¹

The solution doublet lies at the form III band positions, but form II crystallized. towler2007 R5, C12, R9

solid α indomethacin164816791688ethanol solution (gives α)1681nitromethane solution (gives α)16821640165016601670168016901700Raman carbonyl band position, cm⁻¹

The acid–amide band of α near 1648 is absent from both solutions that give α. towler2007 R18, C29

For your drug

  1. Do not choose a crystallization solvent for a target polymorph from solution IR or Raman spectra alone; in this study the forms followed the known supersaturation and temperature rules.
  2. If you use solution spectra, record the concentration of each spectrum, subtract the solvent, include conditions that give each polymorph, and identify the crystallized form on the same solutions by XRPD or Raman. A dimer band in a supersaturated acid solution does not show which form will appear.

towler2007 R1, R2, X5, X6, X7, C26, R17

How it works and key numbers
  1. The starting idea: molecules must cluster before nuclei form, and a cluster motif that matched a polymorph might determine the outcome towler2007 C1, C2.
  2. At low concentration the solute interacts mainly with the solvent; as concentration rises, solute–solute dimers form and gather into solute-rich clusters that exclude solvent (in ethanol the main band moves from 1660 to 1680 cm⁻¹) towler2007 C19, C21, C25.
  3. In ethanol the clusters "appear" γ-like, yet α crystallizes; the authors propose that the few free carbonyls inside dimer clusters "seem" enough to give a pathway to α. None of these steps was measured towler2007 C22, C28, C21, C27.
  4. The extractor's reading: the forms obtained are those the known supersaturation and temperature rules predict (α above a supersaturation of 4 in ethanol, form II below 50 °C), so no role for the solution motif is needed towler2007 R1, R2, X5.
2 of 3informative cases where the solution motif did not match the polymorph that crystallized firsttowler2007 R9, R18, R19
3497/3383 vs 3474/3373 cm⁻¹toluene solution bands sit on form III (3495/3381), not on the form II that crystallizedtowler2007 R9, R5
≈1648 cm⁻¹acid–amide band of solid α, absent from both solutions that give αtowler2007 R18

Connections to other chapters

Evidence: data, conditions and papers

Finding F8. Hydrogen-bond motifs seen in solution did not predict the polymorph that crystallized from solution

In short. IR and Raman spectra of indomethacin and anthranilic acid solutions detected solute–solute contacts, mainly carboxylic acid dimers that form even below saturation, but in two of three informative cases these motifs did not match the polymorph that crystallized first towler2007 R13, R17, R9, R18, R19. Toluene solutions of anthranilic acid had the NH2 bands of form III while form II crystallized, and the acid–amide hydrogen bond that defines α indomethacin was absent from solutions that gave α towler2007 R9, R18, C29. The authors conclude that solution motifs are "not necessarily indicative" of the first polymorph towler2007 C33, C35.

Type. observation in N systems (indomethacin in 2 solvents, anthranilic acid in 5) Strength. the key spectral contrasts are direct and visible in the figures, but the crystallized forms, the concentrations of the spectra and solvent subtraction are not documented, and the conclusion rests on two acids and bulk spectra (towler2007 section 0.8: moderate).

How it was shown

Towler and Taylor (2007) studied two polymorphic carboxylic acids towler2007 A1. Indomethacin was dissolved in ethanol and nitromethane up to a supersaturation c/c* of 5.5, and anthranilic acid in five solvents towler2007 P1, E1, P2. Raman and ATR-IR spectra were set against published solid-state assignments and reference spectra of the forms towler2007 P4, P5, R3, P6.

What the data show

The anthranilic acid Raman spectra differed between the five solvents with no common feature, yet form II was stated to crystallize from every solvent towler2007 R6, R19. Dimer bands grew with concentration in both indomethacin solvents and were present below saturation towler2007 R16, R17, C24.

systemmethod and conditionsresultids
anthranilic acid in five solventsRaman, concentrations that usually give form IIfive different spectra, no common feature; form II from every solventtowler2007 R6, R19, C13
anthranilic acid in tolueneNH2-stretch ATR-IRabout 3497/3383 cm⁻¹ against form III about 3494/3380 and form II about 3471/3370; form II crystallizestowler2007 R9, R5
indomethacin in ethanolIR and Raman up to c/c* 5.5broad band near 1660 cm⁻¹ shifts to 1680; dimer shoulder at 1711 cm⁻¹towler2007 R12, R13
indomethacin in nitromethaneIR at rising concentrationfree carbonyls at 1685 and 1740 cm⁻¹; dimer band at 1713 cm⁻¹ appearstowler2007 R15, C23
indomethacin, α-producing solutionsRaman against solid αno 1650 cm⁻¹ acid–amide band in either solution; solid α has it at about 1648towler2007 R18, C29
Fig. 10 of towler2007: Raman of indomethacin solutions that give α, against solid α
Fig. 10 · towler2007Raman of indomethacin solutions that give α, against solid α

Look at the bottom trace: solid α has a strong band near 1648 cm⁻¹, missing from the ethanol and nitromethane solutions above it, although both give α towler2007 R18.

Why

The authors' starting idea was that molecules must cluster before nuclei form and that the cluster motif might determine the polymorph towler2007 C1, C2. They propose that the few carbonyls freed inside dimer clusters "seem" sufficient to provide a pathway to α towler2007 C22, C28. Cluster size, the free-carbonyl population and nucleation rates were not determined towler2007 C21, C27, C28. (synthesis: the polymorphs obtained are the ones the cited supersaturation and temperature rules predict, so no role for the solution motif is needed towler2007 R1, R2, X5.)

Limits and transfer

Two acids only, under conditions that give their metastable forms towler2007 R19, A1. Spectrum concentrations are not printed and solvent subtraction is not stated towler2007 X6, X7. A dimer band does not by itself indicate which form will appear, because dimers occur in α and γ alike and even below saturation towler2007 C26, R17.

How it was shown

Towler and Taylor (2007) asked whether the hydrogen-bonding motifs that molecules form in solution, as seen by vibrational spectroscopy, foretell which polymorph crystallizes first towler2007 A1. They chose two polymorphic carboxylic acids whose forms differ in hydrogen bonding towler2007 A1. Indomethacin (99%) has a metastable α form, with three molecules in the asymmetric unit of which two form a carboxylic acid dimer and the third an acid–amide hydrogen bond, and a more stable γ form built only of cyclic acid dimers towler2007 M1. Anthranilic acid (≥98%) has form I with a zwitterion and a neutral molecule, metastable form II and high-temperature form III, both of acid dimers towler2007 M2. Indomethacin was dissolved in ethanol (a hydrogen-bond donor and acceptor; solubility 25 mg/mL at 25 °C) and nitromethane (acceptor only; 11 mg/mL) at concentrations up to a supersaturation c/c* of 5.5, with eight and six spectra shown but their concentrations not given towler2007 P1, M3, M4, E1. Anthranilic acid was dissolved in acetonitrile (150 and 52 mg/mL), DMSO (175), ethanol (200), toluene (25) and methanol (25 mg/mL; benzyl alcohol in the Fig. 5 caption), at levels that usually give form II, after heating with excess solid so that each solution was at least saturated at the crystallization temperature towler2007 P2, P3, X2. Raman spectra were recorded with a 785 nm laser at 150 mW (5 s integration, 10 scans) and ATR-IR spectra on a ZnSe accessory at 4 cm⁻¹ resolution with 64 scans; solvent subtraction is not stated towler2007 P4, P5. Solution bands were set against published solid-state assignments of α, γ and amorphous indomethacin from a 1997 library study and against reference spectra of the anthranilic acid forms towler2007 R3, P6. The polymorph crystallized first is stated for each system, but no crystallization procedure or identification method is described towler2007 P7, X5.

What the data show

For anthranilic acid, the Raman spectra in the five solvents differed from one another in band positions and intensities, with no common feature, yet form II was stated to crystallize from every solvent towler2007 R6, R19. The NH2 stretches proved the most useful bands to tell the forms apart: form III near 3495 and 3381 cm⁻¹, form II near 3474 and 3373, form I near 3324 and 3238 towler2007 R5, C12. In toluene the solution doublet sat at about 3497 and 3383 cm⁻¹, on the form III bands (about 3494 and 3380) and not on form II (about 3471 and 3370), at every concentration and temperature measured that leads to form II towler2007 R9. In acetonitrile a carbonyl band at 1670 cm⁻¹, a shoulder at 52 mg/mL and a distinct peak at 150 mg/mL, coincided with a band of solid form II and was assigned to the acid dimer; on the plot it reads near 1678 cm⁻¹ towler2007 R10, C18, X11. For indomethacin, dilute ethanol solutions gave one broad band near 1660 cm⁻¹, assigned to both carbonyls hydrogen bonded to solvent and as broad as the band of amorphous indomethacin towler2007 R12, C19. As concentration rose, a shoulder developed at 1711 cm⁻¹, the asymmetric stretch of an acid dimer, and the main band moved from 1660 to 1680 cm⁻¹ towler2007 R13, C20. In nitromethane, dilute solutions showed free benzoyl (1685) and acid (1740 cm⁻¹) carbonyls, and a dimer band at 1713 cm⁻¹ appeared with concentration towler2007 R15, C23. A band near 920 cm⁻¹, typical of acid dimers, grew with concentration in both solvents, and dimer bands were present in undersaturated as well as supersaturated solutions towler2007 R16, R17, C24. At supersaturations where α crystallizes, solid α showed Raman bands near 1688, 1679 and 1648 cm⁻¹, but neither the ethanol solution (one broad maximum near 1681) nor the nitromethane solution (maximum near 1682) had a band at 1650 cm⁻¹ towler2007 R18.

papersystemmethod and conditionsresultids
towler2007anthranilic acid in five solventsRaman 1750–1310 cm⁻¹, concentrations that usually give form IIfive different spectra, no common feature; form II from every solventR6, R19, C13, C14
towler2007anthranilic acid in toluene, 25 mg/mL and other levelsNH2-stretch ATR-IR≈3497/3383 cm⁻¹ against form III ≈3494/3380 and form II ≈3471/3370; form II crystallizesR9, R5, C17
towler2007anthranilic acid in acetonitrile, 52 and 150 mg/mLcarbonyl IRdimer band (1670 cm⁻¹ in the text, ≈1678 on the plot) grows with concentration and matches a form II bandR10, C18, X8, X11
towler2007indomethacin in ethanolIR and Raman at rising concentration up to c/c* 5.5broad solvent-bonded band ≈1660 cm⁻¹ shifts to 1680; dimer shoulder 1711 cm⁻¹; γ-like by the authors' readingR12, R13, C21, C22
towler2007indomethacin in nitromethaneIR at rising concentrationfree carbonyls 1685 and 1740 cm⁻¹; dimer band 1713 cm⁻¹ appearsR15, C23
towler2007indomethacin, both solvents920 cm⁻¹ band; saturation statedimer band grows with concentration; dimers also below saturationR16, R17, C25
towler2007indomethacin, α-producing solutionsRaman against solid αno 1650 cm⁻¹ acid–amide band in either solution; solid α has it at ≈1648R18, C29

Figure

Fig. 10 of towler2007: Raman of indomethacin solutions that give α, against solid α
Fig. 10 · towler2007Raman of indomethacin solutions that give α, against solid α

Look at the bottom trace: solid α has a strong band near 1648 cm⁻¹, which is missing from the ethanol and nitromethane solutions above it, although both give α towler2007 R18.

Fig. 6 of towler2007: NH2 IR of anthranilic acid in toluene against solid forms II and III
Fig. 6 · towler2007NH2 IR of anthranilic acid in toluene against solid forms II and III

The broad solution doublet sits on the form III bands, not on those of form II, the form that crystallizes towler2007 R9.

Why

The authors' starting idea was that molecules must cluster before nuclei form, and that if the cluster motif matched a polymorph it could determine the outcome ("might") towler2007 C1, C2. In their interpretation, at low concentration the solute interacts mainly with the solvent ("appears"): in ethanol both indomethacin carbonyls are hydrogen bonded to solvent, and for anthranilic acid solute–solvent interactions "appear to dominate" the spectra in every solvent towler2007 C19, C25, C13. As concentration rises, solute–solute dimers form, and the 1660 → 1680 cm⁻¹ shift is linked to molecules gathering into solute-rich clusters that exclude solvent ("We can link") towler2007 C21, C25. In ethanol these clusters "appear" to become more γ-like, yet α crystallizes; the authors propose that the few carbonyls freed inside dimer clusters "seem" sufficient to provide a pathway to α, and that free amide carbonyls in nitromethane "might facilitate" α nucleation towler2007 C22, C28, C27. For anthranilic acid they conclude that solvent "seems" to act on crystal growth (form II morphology) but not on nucleation towler2007 C15. None of these steps was measured: cluster size and composition, the free-carbonyl population and nucleation rates were not determined towler2007 C21, C27, C28. The extractor notes two competing readings: the toluene band position, which did not change with concentration, fits solvated single molecules as well as clusters, and a bulk spectrum cannot exclude a minority population of α-like clusters towler2007 X9, X10. The ethanol "γ-like" reading is also loose, since the solution dimer band (1711–1712 cm⁻¹) lies nearer the amorphous (1710) than the γ position (1718) towler2007 X12. The authors cite competing pictures of the nucleus, including clusters without long-range order and a two-step route through a dense liquid that need not resemble the crystal towler2007 C4. (synthesis: the polymorphs obtained here are the ones the cited supersaturation and temperature rules predict, α above a supersaturation of 4 in ethanol and form II below 50 °C, so the outcome is explained by known conditions without any role for the solution motif towler2007 R1, R2, X5.)

Conditions and limits

Two carboxylic acids only, examined only under conditions that give their metastable forms (α; form II) towler2007 R19, A1. The concentrations, supersaturations and temperatures of the individual spectra are not printed, so which spectra are undersaturated cannot be checked towler2007 X6. Solvent subtraction is not stated, so part of the differences between the anthranilic acid Raman spectra may be solvent bands towler2007 X7. Bulk solution spectra, not time-resolved measurements during nucleation, were used towler2007 P4, P5, X10. Water gave no solute signal for anthranilic acid towler2007 R7, C11.

Counterexamples and conflicts

In acetonitrile the growing dimer band matched form II, the form crystallized towler2007 C18; however, form III contains the same dimer and no form III carbonyl spectrum is shown, so this match is evidence of dimers, not of form II specifically towler2007 X8. A cited earlier study matched dimers in some supersaturated tetrolic acid solutions to the dimers of the polymorph crystallized towler2007 C3. The authors note agreement with a cited 2006 study that solution motifs need not indicate the polymorph towler2007 C33. No other library paper revisited the question.

Transfer check

Do not choose a crystallization solvent for a target polymorph from solution IR or Raman spectra alone; in this study the forms obtained followed the known supersaturation and temperature rules (α above a supersaturation of 4 in ethanol, γ below about 3.5; anthranilic acid form II below 50 °C) towler2007 R1, R2, X5. If solution spectra are used, record the concentration and supersaturation of each spectrum, subtract the solvent, include conditions that give each polymorph, and identify the crystallized form on the same solutions by XRPD or Raman (synthesis: the gaps listed in X5–X7 turned into steps) towler2007 X5, X6, X7. A dimer band in a supersaturated solution of a carboxylic acid drug does not by itself indicate which form will appear, because dimers occur in α and γ alike and even below saturation towler2007 C26, R17.

History

Solid-state IR and Raman assignments of α, γ and amorphous indomethacin came from a 1997 library study of PVP–indomethacin dispersions towler2007 R3. A 2005 study cited in the paper had matched tetrolic acid dimers in solution to the crystallizing polymorph towler2007 C3. Towler and Taylor in 2007 tested the idea on two acids and found the relationship "generally" limited towler2007 C35; no later library paper revisited it.

Topic 2 of 4 · 6 findings · F9–F14

Screening tests built on them

F9

Topic 2Screening tests built on them1 of 6

One heat–cool–heat DSC run sorts a drug into one of three classes by how readily it crystallizes.

Melt the neat crystalline drug, cool it at 20 °C/min and reheat it at 10 °C/min. Of 51 drug-like molecules, 23 crystallized already on cooling (class I), 11 only on reheating (class II) and 17 on neither (class III) baird2010 P1, R1.

moderateModerate: the class counts rest on triplicate DSC runs of 51 molecules and the class I split on two independent methods, but the boundaries are tied to the scan rates and the liquid-N2 cooling rate is unknown baird2010 K1, K2, C21, X12.

51 molecules sorted by one heat–cool–heat cycleClass Icrystallizes on cooling23Class IIcrystallizes only on reheating11Class IIIcrystallizes on neither17

Cooling at 20 °C/min, reheating at 10 °C/min, triplicate runs. baird2010 P1, R1

Class I melts quenched in liquid N2Class I-Astill crystallized15Class I-Bformed a glass8

Birefringence under crossed polarizers and Raman peak broadening gave the same split. baird2010 R3, C10

For your drug

  1. Run the cycle on the neat crystalline drug as published: heat at 10 °C/min to about 10 °C above Tm, hold 3 min, cool at 20 °C/min well below Tg, reheat at 10 °C/min, in triplicate.
  2. If it crystallizes on cooling, quench a 1–2 mg melt in liquid N2 and check for birefringence or broadened Raman bands to separate I-A from I-B. Report the class together with the rates used, because the class can change with them.

baird2010 P1, P5, P6; trasi2012a C7

How it works and key numbers
  1. A glass forms only if the melt is cooled below Tg faster than crystallization can proceed baird2010 C49.
  2. Class I: nucleation and growth are both rapid at overlapping temperatures, so 20 °C/min, and for many even liquid N2, is too slow baird2010 C52.
  3. Class II: cooled fast enough to avoid visible crystals but not fast enough to avoid nuclei. The authors propose that the nuclei remain in the glass and grow on reheating ("most likely") baird2010 C51, C55.
  4. Class III: cooled faster than both limits, which gives a nuclei-free glass, although "it is possible" that a few nuclei form and grow very slowly baird2010 C51, C56.
  5. None of this was measured: nuclei were not detected and rates were not determined. The extractor's reading is that a class shows where the fixed scan rates fall against the drug's own nucleation and growth rates, so it is a rate-specific property (see F10) baird2010 C58, C51; trasi2012a C7.
To feel the numbers. 23 ÷ 51 = 45% of the set crystallized already on cooling; 17 ÷ 51 = 33% made a glass on both steps.
23 / 11 / 17molecules in class I / II / III among the 51baird2010 P1, R1
15 + 8class I molecules that still crystallized (I-A) and that formed a glass (I-B) after liquid-N2 quenchbaird2010 R3
800–1100 °C/mincooling rates at which all eight class I-A molecules tested still crystallizedbaird2013 R1, R3

Connections to other chapters

1 more connection
Evidence: data, conditions and papers

Finding F9. One DSC heat–cool–heat cycle sorts drug molecules into three crystallization-tendency classes

In short. Melting the neat crystalline drug, cooling it at 20 °C/min and reheating at 10 °C/min sorted 51 drug-like molecules into class I (crystallization on cooling, 23), class II (crystallization only on reheating, 11) and class III (neither, 17) baird2010 P1, R1. Quenching the melt in liquid N2 split class I into 15 molecules that still crystallized (I-A) and 8 that formed a glass (I-B), and rapid calorimetry later confirmed that all eight I-A molecules tested crystallize even at 800–1100 °C/min baird2010 R3, C10; baird2013 R1, R3. Later library papers placed about a dozen further drugs with the same test, and assignments agree where a compound recurs raina2015b C12; trasi2020 C7; hiew2022a R3; duong2022a C44; nguyen2023a C59.

Type. observation in N systems (51 molecules; later applied to about a dozen more drugs) Strength. the class counts rest on triplicate DSC runs of 51 molecules (baird2010 K1: strong) and the I-B split on two independent methods (K2: moderate to strong), but the boundaries are tied to the scan rates and the liquid-N2 cooling rate is unknown (baird2010 section 0.8: moderate).

How it was shown

Baird, Van Eerdenbrugh and Taylor (2010) tested 51 neat drug-like molecules by DSC under nitrogen: heat at 10 °C/min to about 10 °C above Tm, hold 3 min, cool at 20 °C/min to −75 °C, reheat at 10 °C/min, in triplicate baird2010 A1, P1. For class I, 1–2 mg melts were plunged into liquid N2 and examined for birefringence and, still immersed, by Raman baird2010 P5, P6. In 2013 class I molecules were re-tested in a rapid heat/cool calorimeter baird2013 P2, A1.

What the data show

Of the 51 molecules, 23 were class I, 11 class II and 17 class III baird2010 R1. After quenching, 15 class I molecules were birefringent and 8 were not baird2010 R3. All eight class I-A molecules still crystallized at 800–1100 °C/min baird2013 R1, R3, R2.

papersystemmethod and conditionsresultids
baird201051 neat moleculesheat–cool–heat, cool 20 °C/min, reheat 10 °C/min23, 11 and 17 in classes I, II and IIIbaird2010 P1, R1
baird201023 class I moleculesliquid-N2 quench; birefringence, Raman8 glassy (I-B), 15 crystalline (I-A)baird2010 P5, P6, R3
baird20138 class I-Arapid heat/cool DSC up to 800–1100 °C/minall crystallized at their fastest coolingbaird2013 P2, R1, R2, R3
lu2016vemurafenibheat–cool–reheat at 10 °C/mincrystallized on cooling from 248.9 °C, about 25 °C below Tmlu2016 R1, C5
trasi2020lumefantrine; artemetherheat–cool–heatlumefantrine class 3, Tg about 18 °C; artemether class 1trasi2020 R5, R6, C7
duong2022adelamanidDSC; liquid-N2 quenchno glass at up to 50 °C/min; class Iduong2022a R3, C44
Fig. 2 of baird2010: Raman spectra after liquid-N2 quench of benzocaine, anthranilic acid and griseofulvin
Fig. 2 · baird2010Raman spectra after liquid-N2 quench of benzocaine, anthranilic acid and griseofulvin

Compare the blue trace (quenched sample in liquid N2) with the black and dashed crystalline references: it overlaps for benzocaine (still crystalline, class I-A) and broadens for griseofulvin (amorphous, class I-B) baird2010 R4, R6.

Why

The authors read the classes on a time–temperature–transformation schematic, not from data baird2010 C51, X5. Class I molecules "clearly" have rapid nucleation and growth at overlapping temperatures baird2010 C52. Class II melts are cooled faster than Rcrit but slower than RNcrit and so, in the authors' reading, keep nuclei that grow on reheating ("most likely") baird2010 C51, C55. Class III melts are cooled faster than both baird2010 C51, C56. Nuclei were not detected baird2010 C58.

Limits and transfer

The liquid-N2 cooling rate was not measured, so the I-A/I-B boundary is tied to that one method baird2010 C21. Run the cycle on the neat crystalline drug exactly as published and report the class with the rates used, because it can change with them baird2010 P1; trasi2012a C7.

How it was shown

Baird, Van Eerdenbrugh and Taylor (2010) asked whether a single DSC heat–cool–heat cycle can classify how easily an organic molecule forms a glass from its melt baird2010 A1. The test set was 51 neat drug-like molecules, described as randomly selected, with melting points from 59 to 272 °C and molecular weights from 121 to 721 g/mol baird2010 A1, M3, M21, M8, M51. Each sample was sealed in a hermetic pan in a TA Q2000 DSC under nitrogen, heated at 10 °C/min to about 10 °C above Tm, held 3 min, cooled at 20 °C/min to −75 °C and reheated at 10 °C/min to just above Tm, in triplicate baird2010 P1. Class I meant crystallization on cooling before the glass transition, class II no crystallization on cooling but crystallization on reheating above Tg, and class III crystallization on neither step baird2010 R1. For class I, 1–2 mg melts on glass slides were plunged into liquid N2 and examined for birefringence under crossed polarizers, and Raman spectra were taken with the sample still immersed so that the laser could not warm and crystallize it baird2010 P5, P6. Baird and co-workers (2013) re-tested eight class I-A and seven class I-B molecules in a prototype rapid heat/cool calorimeter on 30–50 µg single crystals, cooling at 100 and 500 °C/min and at the maximum rate obtainable, 687–1220 °C/min baird2013 P2, A1. Later papers ran the same heat–cool–heat cycle on new drugs, citing the 2010 scheme: six dihydropyridines, lumefantrine and artemether, delamanid, pretomanid and vemurafenib raina2015b R1; trasi2020 R5, R6; duong2022a R3; nguyen2023a C59; lu2016 R1.

What the data show

Of the 51 molecules, 23 were class I, 11 class II and 17 class III baird2010 R1. Four class I molecules (4-biphenyl carboxaldehyde, chlorpropamide, flufenamic acid, tolbutamide) crystallized both on cooling and on reheating, which the authors read as "an indication" that part of each sample had been trapped amorphous baird2010 R2, C9. After liquid-N2 quenching, 15 class I molecules were birefringent and 8 were not, and Raman peak broadening, the sign of amorphous material, appeared for the same 8 baird2010 R3. Quenched griseofulvin, for example, gave a broader carbonyl band reaching about 1725 cm⁻¹, against about 1706–1708 cm⁻¹ for the crystalline references, while benzocaine kept its starting crystal form baird2010 R6, R4. The class I-B glasses crystallized rapidly when reheated above Tg in the Raman experiments baird2010 R15, C24. In the rapid calorimeter all eight class I-A molecules still crystallized at their maximum rates of 800–1100 °C/min; faster cooling only moved the exotherm lower, for phenacetin from about 113 to 107 °C between 100 and 1026 °C/min baird2013 R1, R3, R2. Undercooling at 500 °C/min ranged from 15.1 °C (4-biphenylmethanol) to 90.8 °C (haloperidol) baird2013 R1, C10. The later applications reproduce the 2010 assignments where a compound recurs: felodipine class III and nifedipine class II in both 2010 and 2015, and lumefantrine class III in two papers baird2010 M37, M32; raina2015b C12; trasi2020 C7; hiew2022a R3. Vemurafenib melts crystallized on cooling from 248.9 °C, about 25 °C below Tm, which the authors read as a "relatively poor glass former" lu2016 R1, C5. Delamanid gave no glass at cooling rates up to 50 °C/min, and its liquid-N2 glass devitrified just above Tg, so it was placed in class I with pretomanid duong2022a R3, C44; nguyen2023a C59.

papersystemmethod and conditionsresultids
baird201051 neat moleculesDSC heat–cool–heat, cool 20 °C/min, reheat 10 °C/min, triplicate23 / 11 / 17 in classes I / II / IIIP1, R1, C8
baird201023 class I moleculesliquid-N2 quench of 1–2 mg; PLM and Raman in liquid N28 glassy (I-B), 15 crystalline (I-A); I-B glasses crystallize rapidly on reheatingP5, P6, R3, R15, C10
baird20138 class I-Arapid heat/cool DSC, 30–50 µg, up to 800–1100 °C/minall crystallized at their fastest cooling; exotherm moves lower but persistsP2, R1, R2, R3, C2
lu2016vemurafenibDSC heat–cool–reheat at 10 °C/mincrystallized on cooling from 248.9 °C, about 25 °C below Tm; "relatively poor glass former"R1, C5
raina2015bsix dihydropyridinesDSC heat–cool–reheatfelodipine, cilnidipine, nisoldipine, nitrendipine class III; nifedipine and nimodipine class IIR1, C12
trasi2020lumefantrine; artemetherDSC heat–cool–heatlumefantrine class 3 with Tg about 18 °C; artemether class 1R5, R6, C7
hiew2022alumefantrineDSCclass III; onset Tg 19.7 °CR3
duong2022adelamanidDSC; liquid-N2 quenchno glass at up to 50 °C/min; quenched glass devitrifies just above Tg; class IR3, C44
nguyen2023apretomanid; delamanidDSC and citedboth class I, reduced Tg 0.67 and 0.64C59
moseson2020indomethacin, posaconazole, bicalutamideselection of model drugschosen as class III glass formersC5

Figure

Fig. 2 of baird2010: Raman spectra after liquid-N2 quench of benzocaine, anthranilic acid and griseofulvin
Fig. 2 · baird2010Raman spectra after liquid-N2 quench of benzocaine, anthranilic acid and griseofulvin

Compare the blue trace (quenched sample in liquid N2) with the black and dashed crystalline references: it overlaps for benzocaine (still crystalline, class I-A), shifts to about 1350 cm⁻¹ for anthranilic acid (a new polymorph) and broadens for griseofulvin (amorphous, class I-B) baird2010 R4, R5, R6.

Why

The authors read the classes on a time–temperature–transformation diagram, which they draw as a schematic, not from data baird2010 C51, X5. A glass forms only if the melt is cooled below Tg faster than crystallization can proceed; avoiding any nucleus needs roughly R_{Ncrit} \sim \frac{1}{\tau_{Nmin}}, where τNmin is the shortest nucleation time, which falls as the nucleation rate JN or the sample volume V rises (\tau_N = \frac{1}{J_N V}) baird2010 C49, E21, E22. Because nucleation is faster and acts on a shorter length scale than growth, the rate that avoids detectable crystals (Rcrit) is slower than RNcrit baird2010 C50. Class I molecules "clearly" have rapid nucleation and growth at overlapping temperatures, so 20 °C/min, and for many even liquid N2, is too slow baird2010 C52. Class II melts are cooled faster than Rcrit but slower than RNcrit and so, in the authors' reading, keep nuclei that grow on reheating ("most likely") baird2010 C51, C55. Class III melts are cooled faster than both, giving a nuclei-free glass, although "it is possible" that some nuclei form and grow very slowly baird2010 C51, C56. None of this was measured: nuclei were not detected, and nucleation and growth rates were not determined baird2010 C58, K8. The rapid-calorimetry results were read the same way, as strongly overlapping rate curves for class I-A ("indicate") and less overlap for classes I-B and II baird2013 C33, C35. A 2012 library paper restated the scheme as overlapping (class I), separated (class II) and widely separated or low-rate (class III) nucleation and growth zones trasi2012a C7. (synthesis: each class is a readout of where the screen's fixed cooling and heating rates fall relative to the drug's own nucleation and growth rates, so the class is a rate-specific kinetic property, not a material constant; see F10 baird2010 C51; trasi2012a C7.) The formula behind the argument is on the critical cooling rate to avoid nucleation page.

Conditions and limits

Neat crystalline drugs in hermetic pans at 20 °C/min cooling and 10 °C/min reheating; no polymer, solvent or humidity baird2010 P1, A1. The liquid-N2 cooling rate was not measured, so the I-A/I-B boundary is tied to that one method baird2010 C21. The starting crystal forms are not reported, and niclosamide is listed in the materials without a result baird2010 X11. The Tg values of 16 class I molecules are estimates from molecular weight baird2010 X17. DSC polymorph calls for five compounds differ between two tables baird2010 X2. Class membership near the boundaries depends on the scan rates (see F10) baird2010 X12.

Counterexamples and conflicts

Indoprofen is labelled class I-A in the storage test but class I-B by its own quench data baird2010 X3. Rapid calorimetry moved four class I-B molecules to borderline I-B/II and two class II molecules to borderline II/III baird2013 C13, C31. One later paper calls celecoxib a "slow crystallizer" on the authority of the 2010 scheme, whereas the 2010 paper places it in class II and another later paper lists it as class II mosqueragiraldo2014 R17; baird2010 M26; ilevbare2013c C24 (synthesis: a labelling conflict between papers, not a new measurement). The same molecule can fall in opposite classes by different routes: atenolol is melt class I but spin-coat class III, and dipyridamole is class I from the melt and class III on solvent evaporation vaneerdenbrugh2010a R17; hsieh2012 M5.

Transfer check

Run the cycle on the neat crystalline drug exactly as published: heat at 10 °C/min to about 10 °C above Tm, hold 3 min, cool at 20 °C/min well below Tg, reheat at 10 °C/min, in triplicate baird2010 P1. If it crystallizes on cooling, quench a 1–2 mg melt in liquid N2 and check for birefringence or broadened Raman bands to separate I-A from I-B baird2010 P5, P6. Report the class with the rates used, because the class can change with them trasi2012a C7. A drug inside the evidence is a small neat organic molecule that is thermally stable on melting; a drug that crystallizes even at the fastest cooling available should be treated as class I-A, for which the authors write that knowing Rcrit,cool exceeds 500 °C/min is "probably sufficient" to flag difficulty baird2013 C21.

History

Baird and co-workers introduced the screen in 2010 and offered it as "a useful tool" for early development baird2010 C8, C64. Van Eerdenbrugh and co-workers built a solvent-evaporation analogue the same year vaneerdenbrugh2010a R9. Baird refined the classes with melt viscosity in 2012 and with rapid calorimetry in 2013, which confirmed class I-A and split class I-B baird2012 P2; baird2013 C2, C13. Trasi in 2014 extended the scheme with crystal growth rates into classes IA to IV trasi2014a C17. From 2015 to 2023 the screen was used to classify new drugs and to choose model compounds raina2015b C12; trasi2020 C7; moseson2020 C5; nguyen2023a C59.

F10

Topic 2Screening tests built on them2 of 6

A drug's class can change with the scan rate, and the critical rates differ by orders of magnitude within one class.

The classes were set at fixed rates. Re-tested at other rates, 7 of the 28 class II and III molecules behaved like another class in at least one test, although no class III molecule crystallized on slow cooling baird2010 R7, X12. Within one class, the cooling or heating rate needed to avoid crystallization differed by orders of magnitude baird2013 R5, R9, C13, C22.

moderateModerate: direct DSC data with large, clear spreads, but the number of replicates of the fast runs is not stated and values from two instruments are mixed baird2010 K3; baird2013 K2, K4, X9.

Class II: critical reheating rate, °C/min (log scale)bifonazole12.5celecoxib15tolazamide35droperidol175cinnarizine300clofoctol400dibucaine> 1000acetaminophen> 1910

Slowest reheating that avoided crystallization, eight of the ten class II compounds; 'above' values are drawn at the bound. All ten glasses formed on cooling at 5 °C/min or less. baird2013 R9, C22

4 compounds: still glass at 254-biphenylcarboxaldehyde 687carbamazepine 1220101001000cooling rate, °C/min (log scale)

Class I-B: antipyrin, chlorpropamide, flufenamic acid and tolbutamide formed glasses at 25 °C/min, while the other two still crystallized partly at the fastest rates. baird2013 R5, R9

For your drug

  1. Test the neat drug at the cooling rate of the intended process (for example the cooling after melt extrusion) and reheat at the rates the product will meet, rather than relying on the 20/10 °C/min class.
  2. Treat a drug that changes behaviour between 1 and 20 °C/min cooling, or between 2 and 20 °C/min reheating, as borderline. If a fast-scanning calorimeter is available, measure the critical rates directly, and run replicates, since single crystallization temperatures can scatter by tens of degrees.

baird2010 X12, R7, R8, R9, R12; trasi2012a C7; baird2013 C13, C22

How it works and key numbers
  1. Each class boundary is a fixed rate, 20 °C/min on cooling and 10 °C/min on reheating, so a drug whose own critical rate lies near one of them can move between classes with the protocol (the extractor's reading) baird2010 P1; baird2013 R9.
  2. The authors read the spread within class I-B as compounds that "may have" very different temperature dependencies of nucleation and growth baird2013 C15.
  3. For class II, they consider it "likely" that nucleation and growth overlap little in temperature, so differences in the critical heating rate "can be inferred" as differences in relative nucleation and growth rates; neither rate was measured baird2013 C23.
  4. They attribute the scatter of crystallization temperatures "most likely" to the random nature of nucleation and to unavoidable heterogeneous nucleation in the DSC pans baird2010 C19; baird2013 C18.
To feel the numbers. Within class II, the critical heating rates run from 12.5 to above 1910 °C/min: 1910 ÷ 12.5 = about 150-fold. Within class I-B the rates run from 25 to 1220 °C/min: 1220 ÷ 25 = about 49-fold.
7 of 28class II and III molecules that behaved like another class in at least one testbaird2010 X12
12.5 → above 1910 °C/mincritical heating rate across ten class II glassesbaird2013 R9, C22
25 vs 1220 °C/minclass I-B: four melts still formed glasses at 25, carbamazepine still crystallized partly at 1220baird2013 R5, R9
Evidence: data, conditions and papers

Finding F10. Class membership depends on the scan rates, and critical cooling and heating rates spread over orders of magnitude within a class

In short. Re-tested at other rates, 7 of the 28 class II and III molecules of the 2010 screen behaved like another class in at least one test, while no class III molecule crystallized on slow cooling and every class I-A molecule crystallized at every rate tried baird2010 R7, R8, R9, X12; baird2013 R3. Rapid calorimetry showed that within one class the critical rates differ by orders of magnitude: four class I-B molecules formed glasses at 25 °C/min while others still crystallized at 687–1220 °C/min, and class II glasses needed reheating rates from 12.5 to above 1910 °C/min baird2013 R5, R9, C13, C22. A class is therefore a bin on a continuous rate scale, fixed by the protocol trasi2012a C7.

Type. observation in N systems (28 class II and III molecules re-tested; 25 molecules by rapid heat–cool DSC) Strength. direct DSC data, triplicate in 2010 (baird2010 K3: moderate), with large and clear spreads in 2013 (baird2013 K2: moderate to strong; K4: strong), but the replicate number of the fast runs is not stated and values from two instruments are mixed (baird2013 section 0.8: moderate).

How it was shown

Baird and co-workers (2010) cooled class II and III molecules at 1 °C/min and reheated class II glasses at 20 °C/min and class III glasses at 2 and 10 °C/min baird2010 P4. In 2013 they used a rapid heat/cool calorimeter on 30–50 µg crystals to find the slowest cooling that still gave a glass (Rcrit,cool) and the slowest reheating that avoided crystallization (Rcrit,heat) baird2013 P2, A1, R5, P4.

What the data show

On slow cooling four class II molecules crystallized and no class III molecule did baird2010 R7. Within class II, Rcrit,heat ran from 12.5 to above 1910 °C/min, against Rcrit,cool of 5 °C/min or less for all of them baird2013 R9.

papersystem and conditionsresultids
baird201011 class II and 17 class III; cooling at 1 °C/min, reheating at 20 (class II) and 2 °C/min (class III)4 class II crystallized on slow cooling; bifonazole, celecoxib did not recrystallize at 20 °C/min; nilutamide, nimesulide crystallized at 2 °C/min (2 of 3 runs)baird2010 R7, R8, R9, X12
baird20137 class I-B; maximum, then progressively slower coolingfour glassy down to 25 °C/min; 4-biphenylcarboxaldehyde and carbamazepine partly crystalline at 687 and 1220 °C/min; indoprofen fullybaird2013 R5, R9, C13
baird201310 class II; reheating after 20 °C/min coolingRcrit,heat 12.5 (bifonazole) to above 1910 °C/min (acetaminophen)baird2013 R9, C22
baird20138 class I-A; 100 to 800–1100 °C/minall crystallized at every ratebaird2013 R1, R3
Table 4 of baird2013: critical cooling rate against critical heating rates after slow and after fast cooling
Table 4 · baird2013critical cooling rate against critical heating rates after slow and after fast cooling

Read down the second column, the critical heating rate after 20 °C/min cooling: within class II it runs from 12.5 to above 1910 °C/min, while the first column stays at 5 °C/min or below baird2013 R9, X4.

Why

The authors read the spread within class I-B as compounds that "may have" very different temperature dependencies of nucleation and growth baird2013 C15. For class II they consider it "likely" that nucleation and growth do not overlap much; neither rate was measured baird2013 C23; baird2010 C52. (synthesis: each class boundary is a fixed rate, so a drug whose own critical rate lies near it can move between classes with the protocol baird2010 P1; baird2013 R9.)

Limits and transfer

Values from two instruments are mixed in Table 4 baird2013 X9. The extreme classes held: no class III molecule crystallized on slow cooling, and all eight class I-A molecules crystallized up to 800–1100 °C/min baird2010 R7; baird2013 R1, R3. Test the neat drug at the cooling rate of the intended process and treat a drug that changes behaviour with rate as borderline baird2010 X12, R8; trasi2012a C7.

How it was shown

Because the 20 and 10 °C/min rates of the 2010 screen had been chosen arbitrarily, Baird and co-workers re-tested its classes at other rates baird2010 P4. Class II and III molecules were cooled at 1 °C/min; class II glasses were reheated at 20 °C/min, and class III glasses at 2 °C/min (triplicate) and 10 °C/min baird2010 P4. For the 23 class I molecules the undercooling at crystallization was measured at cooling rates of 5, 10, 15 and 20 °C/min baird2010 P2, R12. In 2013 the same group used a prototype rapid heat/cool calorimeter, calibrated at 10–1700 °C/min, on 30–50 µg crystals of 8 class I-A, 7 class I-B and 10 class II molecules baird2013 P2, A1. Class I-B molecules that did not crystallize at the fastest cooling were cooled at progressively slower rates to find the slowest rate that still gave a glass, the critical cooling rate Rcrit,cool baird2013 P2, R5. Class II glasses made by cooling at 20 °C/min were reheated at 10–200 °C/min in a conventional DSC or at up to about 2000 °C/min in the rapid calorimeter, and the slowest reheating that avoided crystallization was recorded as the critical heating rate Rcrit,heat baird2013 P1, P4. In a separate 2012 study the melts were also sheared at 30 s⁻¹ in a rheometer while cooled stepwise below Tm baird2012 P1.

What the data show

On 1 °C/min cooling, four class II molecules (celecoxib, dibucaine, flurbiprofen, nifedipine) crystallized partly or fully, and none of the 17 class III molecules did baird2010 R7. On 20 °C/min reheating, bifonazole and celecoxib did not recrystallize, while the other nine class II glasses did baird2010 R8. On 2 °C/min reheating, nilutamide and nimesulide (class III) crystallized, each in two of three runs, while the other 15 did not; none of the 17 crystallized at 10 °C/min baird2010 R9. Counted by compound, 7 of the 28 class II and III molecules behaved unlike their class in at least one test, although the authors call the assignments "robust and reasonably independent" of the rates baird2010 X12, C13. Class I crystallization temperatures scattered: at 20 °C/min undercooling ranged from 3.0 °C (caffeine) to 80.8 °C (tolbutamide), antipyrin undercooled by 80.1 °C and the thermally similar 4-biphenylmethanol by 10.1 °C, and for 4-phenylphenol the undercooling was 19.5, 28.4, 27.9 and 23.7 °C at 5, 10, 15 and 20 °C/min, not rising steadily with rate baird2010 R12, C15, X18. In the rapid calorimeter, antipyrin, chlorpropamide, flufenamic acid and tolbutamide (class I-B) did not crystallize at the maximum rate nor at progressively slower rates down to 25 °C/min, while 4-biphenylcarboxaldehyde (687 °C/min) and carbamazepine (1220 °C/min) still crystallized partly and indoprofen fully at the fastest cooling baird2013 R5, R9. Class II glasses needed Rcrit,heat of 12.5 (bifonazole), 15 (celecoxib), 35 (tolazamide), below 50 (D(−) salicin), 175 (droperidol), 300 (cinnarizine), 400 (clofoctol), above 500 (nifedipine), above 1000 (dibucaine) and above 1910 °C/min (acetaminophen), against Rcrit,cool of 5 °C/min or less for all of them baird2013 R9. After quench cooling, bifonazole and celecoxib needed less than 10 °C/min, below the 10 °C/min line that separates classes II and III baird2013 R9, C31. Benzamide and lidocaine crystallized erratically with cooling rate over 5–1004 and 5–839 °C/min as they had over 2–20 °C/min baird2013 R1, R8. Under shear in the rheometer, class I-A melts crystallized at "uniformly higher" temperatures than in DSC baird2012 R5.

papersystemmethod and conditionsresultids
baird201011 class II, 17 class IIIcooling at 1 °C/min; reheating at 20 (class II) and 2 °C/min (class III)4 class II crystallized on slow cooling; bifonazole, celecoxib did not recrystallize at 20 °C/min; nilutamide, nimesulide crystallized at 2 °C/min (2 of 3 runs)R7, R8, R9, X12
baird201023 class Icooling at 5–20 °C/minundercooling at 20 °C/min 3.0 (caffeine) to 80.8 °C (tolbutamide); non-monotonic for 4-phenylphenolR12, C19
baird20137 class I-Brapid calorimetry, maximum then progressively slower coolingfour glassy down to 25 °C/min; 4-biphenylcarboxaldehyde and carbamazepine partly crystalline at 687 and 1220 °C/min; indoprofen fullyR5, R9, C13
baird201310 class IIreheating after 20 °C/min cooling, up to about 2000 °C/minRcrit,heat 12.5 (bifonazole) to above 1910 °C/min (acetaminophen)R9, C22
baird2013bifonazole, celecoxibreheating after quench coolingRcrit,heat below 10 °C/min, borderline class II/IIIR9, C31
baird20138 class I-A100 °C/min to 800–1100 °C/minall crystallized at every rateR1, R3
baird2012class I-Arheometer, 30 s⁻¹ shearcrystallization temperatures uniformly higher than in DSCR5, C8
trasi2012areview of the classes—"the classification category of a given compound can be altered" by the ratesC7

Figure

Table 4 of baird2013: critical cooling rate against critical heating rates after slow and after fast cooling
Table 4 · baird2013critical cooling rate against critical heating rates after slow and after fast cooling

Read down the second column, which is the critical heating rate after 20 °C/min cooling although its printed head says Rcrit,cool: within class II it runs from 12.5 to above 1910 °C/min, while the first column stays at 5 °C/min or below baird2013 R9, X4.

Why

The authors read the spread within class I-B as compounds that "may have" very different temperature dependencies of nucleation and growth baird2013 C15. For class II, which did not crystallize on slow cooling, they consider it "likely" that nucleation and growth do not overlap much in temperature, so that differences in Rcrit,heat "can be inferred" as differences in relative nucleation and growth rates spanning orders of magnitude baird2013 C23. Nucleation and growth rates were not measured as functions of temperature in either paper baird2013 C23; baird2010 C52. They attribute the scatter of crystallization temperatures "most likely" to the stochastic nature of nucleation and to unavoidable heterogeneous nucleation in the DSC pans baird2010 C19; baird2013 C18. The higher crystallization temperatures under shear are put down to heterogeneous nucleation at the upper rheometer plate baird2012 C8. (synthesis: each class boundary is a fixed rate, 20 °C/min on cooling and 10 °C/min on reheating, so any drug whose own critical rate lies near one of these values can move between classes with the protocol; the 2013 critical rates replace the three bins by a continuous measure baird2010 P1; baird2013 R9.)

Conditions and limits

Neat drugs in DSC pans; conventional DSC used 3–5 mg samples in triplicate, the rapid calorimeter 30–50 µg crystals with an unstated number of replicates baird2013 P1, P2, X9. Table 4 of the 2013 paper mixes values from two instruments without stating which gave each, and heating rates of 1900–2000 °C/min lie outside the calibrated range baird2013 X9, X10. The quench rate before reheating is printed for only three compounds baird2013 X5. The 2010 tests used 1 °C/min cooling but no 2 °C/min cooling, so the statement that classes II and III form glasses at 1–2 °C/min holds for class III and 7 of 11 class II molecules only baird2010 X13.

Counterexamples and conflicts

The extreme classes held: no class III molecule crystallized on cooling at 1 °C/min or reheating at 10 °C/min, and all eight class I-A molecules crystallized up to 800–1100 °C/min baird2010 R7, R9; baird2013 R1, R3. Rates above 1000 °C/min were reached for only four of the eight class I-A molecules, so "above 1000 °C/min" is a bound per compound baird2013 X6. The Table 4 value for clofoctol (400 °C/min) conflicts with the text, which names 500 °C/min as the rate that removes the endotherm baird2013 X14.

Transfer check

Test the neat drug at the cooling rate of the intended process (for example the cooling after melt extrusion) and reheat at the rates the product will meet, rather than relying on the 20/10 °C/min class baird2010 X12; trasi2012a C7. Treat a drug that changes behaviour between 1 and 20 °C/min cooling, or between 2 and 20 °C/min reheating, as borderline baird2010 R7, R8, R9. Where a fast-scanning calorimeter is available, measure Rcrit,cool and Rcrit,heat directly, because these spread over orders of magnitude within one class baird2013 C13, C22. Run replicates, since single crystallization temperatures can scatter by tens of degrees baird2010 R12.

History

Baird in 2010 found most assignments robust while recording the deviations baird2010 C13, R7, R8, R9. Trasi in 2012 stated that the class of a compound can be altered by the heating and cooling rates trasi2012a C7. Baird in 2013 measured the spread of critical rates within classes and proposed borderline classes I-B/II and II/III baird2013 C13, C22, C31.

F11

Topic 2Screening tests built on them3 of 6

Slow DSC scans cannot reliably predict how fast a drug must cool to stay amorphous; wider rate ranges help.

The critical cooling rate (the slowest cooling that avoids crystals) can be extrapolated from how far a melt undercools at several cooling rates. With rates of 5–20 °C/min this failed for most class I molecules: 12 of 19 fits had r² below 0.55 baird2010 R13, C20. With rates up to about 800–1100 °C/min, six compounds gave r² of 0.89–0.99 baird2013 R1, R7, R8.

moderateModerate: the failure shows both as poor fits and as a contradiction with quench results, but the improvement rests on six fits, and three printed table values are misprinted or cannot be reproduced baird2010 K4; baird2013 K3, X2, X3, X13.

19 fits of the critical cooling rate over 5–20 °C/minr² 0.74–0.99usable7 fitsr² below 0.55failed12 fits

Undercooling often did not rise steadily with cooling rate, and crystallization temperatures scattered between runs. baird2010 R13, C18

haloperidol 20 · crystallizedphenacetin 64 · crystallizedcarbamazepine 118 · glassgriseofulvin 588 · glass101001000estimated critical cooling rate, K/min (log scale)

A low estimate should indicate easy glass formation; the liquid-N2 quench test showed the reverse. baird2010 R13, R3, C21

For your drug

  1. Answer the practical question directly with a quench test or a fast-scanning DSC run, rather than extrapolating from a conventional DSC.
  2. If you do extrapolate, use cooling rates spanning several orders of magnitude, check that undercooling rises steadily with rate across replicates, and compare the estimate with the outcome at the fastest rate available. A drug that crystallizes at 500 °C/min or faster falls in the "difficult to make and keep amorphous" group.

baird2013 C21, C17, C20; baird2010 C20, C22

How it works and key numbers
  1. The shortcut comes from inorganic glasses: measure the undercooling \Delta T = T_m - T_{cryst,max} at several cooling rates, fit \ln(\text{rate}) against 1/\Delta T^2, and read the critical rate from the intercept A as R_{crit} = \exp(A) baird2010 E1, E2, E3.
  2. Both methods assume that a higher cooling rate always lowers the crystallization temperature, so that undercooling grows regularly baird2013 C17; baird2010 R31.
  3. With organic drugs, undercooling often did not rise steadily with rate and crystallization temperatures scattered between runs; the authors put this down "most likely" to the random nature of nucleation and to heterogeneous nucleation in DSC pans baird2010 C18, C19; baird2013 C18.
  4. A window of 5–20 °C/min lies far below the critical rate of class I molecules, which crystallize even in liquid N2, so the extrapolation is long; for inorganic glasses accurate predictions need rates at least two orders of magnitude below Rcrit baird2010 C22, R3, R31.
  5. A wider range "seems to improve" the fits: atenolol, benzocaine and felbinac went from r² below 0.55 to r² of 0.93 or more. Benzamide, lidocaine and indoprofen still failed baird2013 R8, C20, R7, C16.
To feel the numbers. The haloperidol intercept of 6.5 gives e^{6.5} \approx 665 K/min, not the printed 6.6 × 10³. That is below the 913 °C/min at which haloperidol still crystallized, so even the wide-range fit underestimates for this compound.
12 of 19fits over 5–20 °C/min with r² below 0.55baird2010 R13
0.89–0.99r² of six fits over cooling rates up to 800–1100 °C/minbaird2013 R7
500 °C/mincooling rate above which the authors say a drug is flagged as hard to make and keep amorphous ("probably sufficient")baird2013 C21
Evidence: data, conditions and papers

Finding F11. Critical cooling rates extrapolated from 5–20 °C/min DSC data are unreliable for organic molecules; a much wider rate range helps

In short. The Barandiaran–Colmenero and Cabral extrapolations, applied to undercooling measured at 5–20 °C/min, failed for most class I molecules (12 of 19 fits with r^2 < 0.55), and the usable estimates contradicted the liquid-N2 quench results baird2010 R13, R3, C20, C21. Extending the cooling range to about 800–1100 °C/min by rapid calorimetry gave r² 0.89–0.99 for six compounds, including three whose narrow-range fits had failed, but the method still failed where crystallization temperatures scattered, and one printed estimate is likely ten times too high baird2013 R1, R7, R8, C16, C20, X2. The authors conclude that knowing Rcrit,cool exceeds 500 °C/min is "probably sufficient" to flag a drug that will be hard to make and keep amorphous baird2013 C21.

Type. method comparison Strength. the failure is shown both by poor fits and by contradiction with quench outcomes (baird2010 K4: moderate), while the improvement rests on six fits with three misprinted or non-reproducible table values (baird2013 K3: moderate).

How it was shown

Rcrit can be extrapolated by regressing \ln(\text{rate}) on 1/\Delta T^2 of the undercooling \Delta T = T_m - T_{cryst,max} at several cooling rates, taking R_{crit} = \exp(A) (Barandiaran–Colmenero) or, with Cabral's correction, \exp\left(A^* - \frac{B^*}{T_m^2}\right) baird2010 E1, E2, E3. Baird and co-workers (2010) used 5–20 °C/min on class I molecules and checked the estimates against liquid-N2 quenching baird2010 P2, P9, P6, R3. In 2013 they refitted eight class I-A molecules and indoprofen up to 800–1100 °C/min baird2013 P3, R1.

What the data show

Of the 19 fits in 2010, seven had r² 0.74–0.99 and twelve r^2 < 0.55, because undercooling did not rise steadily with cooling rate baird2010 R13, C18. Over the wider 2013 range, six compounds gave r² 0.89–0.99 baird2013 R7.

papersystemresultids
baird201019 class I molecules, fits over 5–20 °C/min7 fits with r² 0.74–0.99, 12 with r^2 < 0.55baird2010 R13, C18
baird2010haloperidol, phenacetin; carbamazepine, griseofulvinhaloperidol (20 K/min) and phenacetin (64) crystallized in liquid N2; carbamazepine (118) and griseofulvin (588) vitrifiedbaird2010 R13, R3, C21
baird20138 class I-A and indoprofen, fits over 5 to about 800–1100 °C/minr² 0.89–0.99 for six; Rcrit,cool 2.7–8.1 × 10³ K/min as printed; benzamide, lidocaine, indoprofen r^2 < 0.55baird2013 R7, C16, C19
baird2013atenolol, benzocaine, felbinacr^2 < 0.55 over 2–20 °C/min, r^2 \geq 0.93 over the wide rangebaird2013 R8, C20
baird2013haloperidol6.6 × 10³ K/min printed; the printed intercept implies about 6.6 × 10² K/minbaird2013 X2
Table 3 of baird2013: BC and Cabral estimates of the critical cooling rate over a wide range of cooling rates
Table 3 · baird2013BC and Cabral estimates of the critical cooling rate over a wide range of cooling rates

Three compounds fail with r^2 < 0.55, and the haloperidol value should be read as about 6.6 × 10² rather than 6.6 × 10³ K/min baird2013 R7, X2.

Why

Both methods assume that a higher cooling rate always lowers the crystallization temperature, so that undercooling grows regularly toward a limit baird2013 C17; baird2010 R31. The authors attribute the failures to scattered crystallization temperatures, put down "most likely" to the stochastic nature of nucleation and to heterogeneous nucleation in DSC pans baird2010 C18, C19; baird2013 C18.

Limits and transfer

Only neat class I molecules were fitted, since class II and III molecules do not crystallize on cooling at these rates baird2010 P9. The liquid-N2 cooling rate was not measured baird2010 C21. Answer the practical question directly with a quench test or a fast-scanning DSC run rather than extrapolating from a conventional DSC baird2013 C21; baird2010 C20.

How it was shown

The critical cooling rate Rcrit, the slowest cooling that avoids detectable crystallization, is the usual measure of glass-forming ability baird2010 C50, E2. Inorganic-glass science offers a shortcut: measure the undercooling \Delta T = T_m - T_{cryst,max} at several cooling rates, regress \ln(\text{rate}) on 1/\Delta T^2, and take Rcrit from the intercept A as R_{crit} = \exp(A) (Barandiaran–Colmenero, BC) or, with Cabral's correction, as R_{crit} = \exp\left(A^* - \frac{B^*}{T_m^2}\right) baird2010 E1, E2, E3. These methods had given reasonable values for inorganic glasses but had not been tested on organic molecules baird2010 C22, R31. Baird and co-workers (2010) applied both to the 23 class I molecules of their DSC screen, using undercooling at 5, 10, 15 and 20 °C/min; the four molecules that crystallized only partly on cooling were left out, leaving 19 fits baird2010 P2, P9, R13. Where crystallization was so fast that it heated the thermocouple, the peak temperature was estimated from the time to maximum heat flow baird2010 P2. The estimates were checked against the outcome of quenching 1–2 mg melts in liquid N2 baird2010 P6, R3. In 2013 the same group refitted eight class I-A molecules and indoprofen with undercooling measured from 5 °C/min (conventional DSC) to the maximum rate of a rapid heat/cool calorimeter, 800–1100 °C/min baird2013 P3, R1.

What the data show

Of the 19 fits in 2010, seven had r² 0.74–0.99 and twelve had r^2 < 0.55, because undercooling often did not rise steadily with cooling rate and crystallization temperatures scattered between runs baird2010 R13, C18. Where the fits worked, Rcrit ran from 20 K/min (haloperidol) to 3.28 × 10⁴ K/min by BC and 3.26 × 10⁴ K/min by Cabral (4-biphenylmethanol) baird2010 R13. The estimates contradicted the quench test: haloperidol (20 K/min) and phenacetin (64 K/min) still crystallized in liquid N2, whereas carbamazepine (118 K/min by BC, 117 by Cabral) and griseofulvin (588 and 579 K/min) were quenched amorphous, the reverse of the order the estimates predict baird2010 R13, R3, C21. The authors conclude that the approach "may be of limited utility" for organic molecules and that rates spanning several orders of magnitude "may be needed", noting that for inorganic glasses accurate predictions need cooling rates at least two orders of magnitude below Rcrit baird2010 C20, C22, R31. Over the wider 2013 range, the fits gave r² 0.89–0.99 for six compounds, with printed BC estimates of 8.1 × 10³ (4-biphenylmethanol), 3.2 × 10³ (atenolol), 6.2 × 10³ (benzocaine), 3.0 × 10³ (felbinac), 6.6 × 10³ (haloperidol) and 2.7 × 10³ K/min (phenacetin) baird2013 R7. Atenolol, benzocaine and felbinac, which had given r^2 < 0.55 over 2–20 °C/min, now gave r^2 \geq 0.93, so a wider range "seems to improve" the fits baird2013 R8, C20. Benzamide, lidocaine and indoprofen still failed (r^2 < 0.55), because their undercooling rose and fell with rate baird2013 R7, C16. Three printed values do not follow from their own inputs: haloperidol's intercept 6.5 gives \exp(6.5) \approx 6.6 \times 10^2 K/min, not 6.6 × 10³; the 4-biphenylmethanol intercept lacks a digit; and the felbinac fit cannot be reproduced baird2013 X2, X3, X13.

papersystemmethod and conditionsresultids
baird201019 class I moleculesBC and Cabral fits over 5–20 °C/min7 fits with r² 0.74–0.99, 12 with r^2 < 0.55; estimates 20 to 3.28 × 10⁴ K/minR13, C18
baird2010haloperidol, phenacetin; carbamazepine, griseofulvinestimates against liquid-N2 quenchhaloperidol (20 K/min) and phenacetin (64) crystallized; carbamazepine (118) and griseofulvin (588) vitrifiedR13, R3, C21
baird2010inorganic glasses (cited)literatureaccurate predictions need rates at least two orders of magnitude below RcritR31, C22
baird20138 class I-A and indoprofenfits over 5 to about 800–1100 °C/minr² 0.89–0.99 for six; Rcrit,cool 2.7–8.1 × 10³ K/min as printed; benzamide, lidocaine, indoprofen r^2 < 0.55R7, C16, C19
baird2013atenolol, benzocaine, felbinacnarrow against wide rate ranger^2 < 0.55 over 2–20 °C/min, r^2 \geq 0.93 over the wide rangeR8, C20
baird2013haloperidolprinted against recomputed estimate6.6 × 10³ K/min printed; the printed intercept implies about 6.6 × 10² K/minX2

Figure

Table 3 of baird2013: BC and Cabral estimates of the critical cooling rate over a wide range of cooling rates
Table 3 · baird2013BC and Cabral estimates of the critical cooling rate over a wide range of cooling rates

Each row gives the intercept, slope, r² and the two Rcrit estimates; three compounds fail with r^2 < 0.55, and the haloperidol value should be read as about 6.6 × 10² rather than 6.6 × 10³ K/min baird2013 R7, X2.

Why

Both methods assume that a higher cooling rate always lowers the crystallization temperature, so that undercooling grows in a regular way toward a limit as the rate approaches Rcrit baird2013 C17; baird2010 R31. The authors attribute the failures to undercooling that did not rise consistently with rate and to scattered crystallization temperatures, which they put down "most likely" to the stochastic nature of nucleation and to heterogeneous nucleation in DSC pans baird2010 C18, C19; baird2013 C18. A rate window of 5–20 °C/min lies far below the Rcrit of class I molecules, which crystallize even when quenched in liquid N2, so the extrapolation is long baird2010 C22, R3. The 2013 estimates, except haloperidol's, are much larger than the maximum rates reached and so "may be reasonable estimates" baird2013 C19. (synthesis: with the corrected haloperidol value of about 6.6 × 10² K/min, the extrapolated Rcrit lies below the 913 °C/min at which haloperidol still crystallized, so even the wide-range fit underestimates for this compound; this repeats in milder form the 2010 contradiction with the quench test baird2013 X2, R1.) The formulas are on the Barandiaran-Colmenero critical cooling rate and Cabral critical cooling rate pages.

Conditions and limits

Neat class I molecules only; class II and III molecules do not crystallize on cooling at these rates and cannot be fitted this way baird2010 P9. The liquid-N2 cooling rate used as the check was not measured baird2010 C21. In 2013 the points used per fit and the definition of Tcryst are not fully stated, and the indoprofen undercooling row duplicates phenacetin's, so indoprofen's own data are missing baird2013 X16, X1. Replicate numbers of the fast runs are not stated baird2013 X9.

Counterexamples and conflicts

None found in the library supporting the narrow-range method for organic molecules. Within the 2010 data some fits were good (carbamazepine r² 0.99), so a good fit did not guarantee a correct ranking against the quench test baird2010 R13, C21. The 2013 paper partly revises the 2010 verdict: the method works for some compounds when the rate range is wide baird2013 C20.

Transfer check

For a new drug, answer the practical question directly with a quench test or a fast-scanning DSC run rather than extrapolating from a conventional DSC baird2013 C21; baird2010 C20. If an extrapolation is attempted, use cooling rates spanning several orders of magnitude, check that undercooling rises steadily with rate across replicates, and compare the estimate with the outcome at the fastest rate available baird2010 C22; baird2013 C17, C20. A drug that crystallizes at 500 °C/min or faster falls inside the "difficult to make and keep amorphous" group regardless of the exact estimate baird2013 C21.

History

The BC (1981) and Cabral (2003) methods came from inorganic glass science and are cited baird2010 E1, E3. Baird in 2010 found them of "limited utility" for organic molecules over 5–20 °C/min baird2010 C20. Baird in 2013 found that a much wider rate range improves the fits but still fails for compounds with erratic crystallization baird2013 C20, C16.

F12

Topic 2Screening tests built on them4 of 6

Simple temperature ratios do not tell you whether a drug will crystallize.

The reduced glass transition temperature T_{rg} = T_g/T_m, glass-stability parameters and fragility come from inorganic glass science. Tested on the drug molecules of the 2010 screen, none of them separated good from poor glass formers baird2010 C38; baird2012 C39.

moderateThe failures are visible in the tabulated values, but no statistical test is named and 16 class I Tg values are estimates baird2010 K5, X17.

inorganic rule: higher Tg/Tm, better glassClass Icrystallizes on coolingmean 0.69 ± 0.02Class IIcrystallizes on reheatingrange 0.65–0.76Class IIIstays amorphousmean 0.73 ± 0.04range 0.65–0.810.60.650.70.750.80.85reduced glass transition temperature, Tg/Tm

The class means lie in the expected order, but the difference was not significant and the ranges overlap. baird2010 R14, R18, C37

Named best glass formers by the parameters
bifonazolecelecoxibcinnarizinedroperidol
Actually crystallized on 1 °C/min cooling
celecoxibdibucaineflurbiprofennifedipine

Class II molecules; only celecoxib appears on both lists. baird2010 C27, R7, C28

For your drug

  1. Do not screen candidates by Tg/Tm, glass-stability parameters or fragility alone.
  2. Run the DSC heat–cool–heat test, measure a seeded growth rate, and if possible the melt viscosity at Tm and its rise over the first 40 °C of undercooling. If you report Tg/Tm, measure Tg rather than estimating it from molecular weight.

baird2010 C38, C64, X17; baird2012 C36, C43, C45

How it works and key numbers
  1. T_g/T_m assumes that every liquid has the same viscosity at Tg, so a higher ratio would mean a more viscous liquid over the undercooled range baird2010 C1.
  2. The drug data contradict this assumption: compounds with almost the same Tm − Tg showed very different rises in viscosity on cooling baird2012 C36.
  3. All 28 melts fitted with the VTF equation were fragile (D 4.3–8.7) whatever their class, so fragility cannot separate them either baird2012 R12, C39.
Same Tm − Tg (105–108 °C): viscosity rise over 40 °C of coolingchlorpropamideclass I-B · 108 °C10×celecoxibclass II · 105 °C20×ketoconazoleclass III · 105 °C36×

baird2012 M9, M14, M26, R3

51drug molecules whose Tg/Tm was calculated in the 2010 screenbaird2010 E4, P8
18 → 731spread of the fragility index m, with no pattern by classbaird2010 R18, C31
"may not be ideal"the authors' verdict on these parameters for organic moleculesbaird2010 C38

Connections to other chapters

Evidence: data, conditions and papers

Finding F12. Reduced Tg, glass-stability parameters and kinetic fragility do not rank organic molecules by crystallization tendency

In short. Mean Tg/Tm was 0.69 ± 0.02 for class I against 0.73 ± 0.04 for class III of the 2010 screen, in the expected order but not significantly different, and the ranges overlapped baird2010 R14, C37. Glass-stability parameters built from crystallization on reheating picked the wrong class II molecules as the best glass formers baird2010 C27, C28. Kinetic fragility did not follow the classes either: the fragility index m ran from 18 to 731 without a class pattern, and every one of 28 melts fitted with the VTF equation was fragile (D 4.3–8.7) whatever its class baird2010 R18, C31; baird2012 R12, C39. Compounds with almost the same Tm − Tg fell in classes I-B, II and III baird2012 M9, M14, M26, C36.

Type. observation in N systems (51 molecules; 28 fitted viscosity curves) Strength. the failures are visible directly in the tabulated values (baird2010 K5: moderate; baird2012 K4: moderate, K5: moderate to strong), but no statistical test is named and 16 class I Tg values are estimates (baird2010 X17).

How it was shown

Baird and co-workers (2010) calculated T_{rg} = \frac{T_g}{T_m} for the 51 molecules of their DSC screen, glass-stability parameters for 11 class II and 3 class I-B molecules, and fragility m from the rate dependence of Tg baird2010 E4, P8, P11, P3, P10. In 2012 they measured the melt viscosity of 34 molecules and fitted the VTF equation to 28 curves to obtain the strength parameter D baird2012 P1, P5, P6.

What the data show

Trg did not separate the classes, the stability parameters pointed the wrong way inside class II, and all fitted melts were fragile whatever their class baird2010 R14, C37, C27, C28; baird2012 R12, C39.

paperpredictorresultids
baird2010T_{rg} = \frac{T_g}{T_m}, 51 moleculesI-A 0.69, I-B 0.7; class I 0.69 ± 0.02 against class III 0.73 ± 0.04, not significant; class II 0.65–0.76, class III 0.65–0.81baird2010 R14, R18, C23, C37
baird2010glass-stability parameters, 11 class IIpredicted best: bifonazole, celecoxib, cinnarizine, droperidol; crystallized at 1 °C/min: celecoxib, dibucaine, flurbiprofen, nifedipinebaird2010 C27, C28, R7
baird2010fragility m18 (tolazamide) to 731 (itraconazole); no correlation with glass-stability parametersbaird2010 R18, C31
baird2012VTF strength D, 28 compounds4.3–8.7 in all classes; one band on the Tg-scaled plotbaird2012 R12, R13, C39, C42
baird2012chlorpropamide, celecoxib, ketoconazolesimilar Tm − Tg (108, 105, 105 °C); viscosity ratio at 40 °C undercooling about 10, 20 and 36; classes I-B, II, IIIbaird2012 M9, M14, M26, R3
Fig. 5 of baird2012: all measured melt viscosities on a Tg-scaled (Angell) plot with VTF curves for D = 100 and D = 5
Fig. 5 · baird2012all measured melt viscosities on a Tg-scaled (Angell) plot with VTF curves for D = 100 and D = 5

Points of every class sit in one band around the fragile D = 5 curve; the class colours do not separate, so fragility on this scale carries no class information baird2012 R13, C42.

Why

The reduced Tg rests on the assumption that all liquids have the same viscosity at Tg baird2010 C1. The 2012 data contradict it: compounds with similar viscosity at Tm and similar Tm − Tg can differ in how fast viscosity rises on cooling, so Tg/Tm "should not be considered a reliable predictor" baird2012 C36. Organic liquids are mostly fragile, so Angell plots "give minimal information" on crystallization tendency baird2012 C40, C43.

Limits and transfer

No statistical test is named, and 16 class I Tg values are estimates from molecular weight baird2010 R14, X17. The fragility category depends on whether m or D is used baird2010 X14. Do not screen candidates by Tg/Tm, glass-stability parameters or fragility alone; the heat–cool–heat test, a seeded growth rate and the melt viscosity at Tm carried more information in these data sets baird2010 C38; baird2012 C45; baird2010 C64.

How it was shown

Inorganic-glass science offers simple predictors of glass-forming ability: the reduced glass transition temperature T_{rg} = \frac{T_g}{T_m} (in kelvin), for which a higher value is taken to mean a more viscous liquid between Tg and Tm and so more resistance to crystallization, with a typical value of 0.66 proposed baird2010 E4, C1, R29. Glass-stability parameters (KLL, KH, KW, KSP, K1, K2 and the reduced temperature Tred) combine Tg, the crystallization temperatures on reheating and Tm, and had risen as the measured Rcrit fell in inorganic glasses baird2010 E5, E11, C2, R28. Good glass formers were also expected to be "strong" liquids, whose mobility changes little near Tg baird2010 C3. Baird and co-workers (2010) calculated Trg for all 51 molecules of their DSC screen, using measured or literature Tg and, for 16 class I molecules that crystallized too fast to show a Tg, a value estimated from a regression of Trg on molecular weight baird2010 E4, E27, P8. Glass-stability parameters were calculated for the 11 class II molecules and three class I-B molecules baird2010 P11. Fragility m, strength D and the Kauzmann temperature came from the shift of Tg with heating and cooling rate (5–20 °C/min) and from the heat-capacity step, for classes II and III baird2010 P3, P10. Baird and co-workers (2012) measured the steady-shear viscosity of 34 of the same molecules from Tm downward at 30 s⁻¹, fitted the VTF equation \eta = \eta_0 \exp\left(\frac{D\,T_0}{T - T_0}\right) to 28 curves of classes I-B, II and III, and plotted all data on a Tg-scaled (Angell) plot baird2012 P1, P5, P6, E4.

What the data show

Mean Trg was 0.69 (±0.01) for class I-A and 0.7 (±0.03) for class I-B, not different, so Trg did not predict which class I molecules could be quenched to a glass baird2010 R14, C23. Class I as a whole averaged 0.69 ± 0.02 against 0.73 ± 0.04 for class III, the expected order but not a significant difference, so Trg is "clearly not an infallible predictor" baird2010 R14, C37. Trg spanned 0.65–0.76 in class II and 0.65–0.81 in class III, and the two class III molecules that crystallized on slow reheating did not have lower values (nilutamide 0.72, nimesulide 0.70) baird2010 R18, C36. Within class II, Tred and the glass-stability parameters pointed to bifonazole, celecoxib, cinnarizine and droperidol as the best glass formers, yet celecoxib, dibucaine, flurbiprofen and nifedipine were the ones that crystallized on cooling at 1 °C/min baird2010 C27, R7, C28. Tred after 20 °C/min cooling ran from 0.33 (acetaminophen) to 0.72 (celecoxib), but celecoxib also had the lowest KSP (1.464) and K2 (1.118), so the ranking depended on the parameter chosen baird2010 R16, R17, X15. The fragility index m ran from 18 (tolazamide) to 731 (itraconazole) across classes II and III, and neither fragility nor strength correlated with the glass-stability parameters baird2010 R18, C31. From viscosity, all 28 fitted compounds had D between 4.3 and 8.7, below the fragile limit of 10, with class III (4.3–6.3) no less fragile than class I-B (4.5–7.3) or class II (4.4–8.7) baird2012 R12, C39. On the Angell plot the data of all four classes lie in one band near the D = 5 curve, far from the strong D = 100 line baird2012 R13, C42. Chlorpropamide (class I-B), celecoxib (class II) and ketoconazole (class III) have Tm − Tg of 108, 105 and 105 °C, yet their viscosity rose about 10-, 20- and 36-fold over 40 °C of undercooling baird2012 M9, M14, M26, R3. Class means of Tm − Tg (123, 119, 126 and 108 °C for classes I-A to III) did not differ significantly baird2012 R10. Later, pretomanid and delamanid were placed in class I with reduced Tg 0.67 and 0.64 nguyen2023a C59.

papersystemmethod and conditionsresultids
baird201051 moleculesTg/Tm in kelvin; Tg of 16 class I molecules from MWI-A 0.69, I-B 0.7; class I 0.69 ± 0.02 against class III 0.73 ± 0.04, not significant; class II 0.65–0.76, class III 0.65–0.81R14, R18, C23, C37, X17
baird201011 class IIKLL, KH, KW, KSP, K1, K2, Tredpredicted best: bifonazole, celecoxib, cinnarizine, droperidol; observed to crystallize at 1 °C/min: celecoxib, dibucaine, flurbiprofen, nifedipineC27, C28, R7, X15
baird2010classes II and IIIm from the rate dependence of Tgm 18 (tolazamide) to 731 (itraconazole); no correlation with glass-stability parametersR18, C31, X14
baird201228 compoundsVTF fit of melt viscosityD 4.3–8.7 in all classes; one band on the Tg-scaled plotR12, R13, C39, C42, C43
baird2012chlorpropamide, celecoxib, ketoconazolesimilar Tm − Tg (108, 105, 105 °C)viscosity ratio at 40 °C undercooling about 10, 20 and 36; classes I-B, II, IIIM9, M14, M26, R3, C36
baird201234 compoundsclass means of Tm − Tg123, 119, 126, 108 °C, not significantly differentR10
nguyen2023apretomanid, delamanidTg/Tm0.67 and 0.64, both class IC59

Figure

Fig. 5 of baird2012: all measured melt viscosities on a Tg-scaled (Angell) plot with VTF curves for D = 100 and D = 5
Fig. 5 · baird2012all measured melt viscosities on a Tg-scaled (Angell) plot with VTF curves for D = 100 and D = 5

Points of every class sit in one band around the fragile D = 5 curve; the class colours do not separate, so fragility on this scale carries no class information baird2012 R13, C42.

Fig. 1 of baird2010: Trg against molecular weight with the regression used to estimate 16 Tg values
Fig. 1 · baird2010Trg against molecular weight with the regression used to estimate 16 Tg values

Trg rises only weakly with molecular weight (R² 0.44), and the 16 estimated class I values lie on this line, so their Trg is a function of molecular weight alone baird2010 R19, X17.

Why

The reduced Tg rests on the assumption that all liquids have the same viscosity at Tg, so that a higher Tg/Tm means a more viscous liquid over the undercooled range baird2010 C1. The 2012 data contradict that premise: compounds with similar viscosity at Tm and similar Tm − Tg can differ in how fast viscosity rises on cooling, as chlorpropamide and ketoconazole do, so Tg/Tm "should not be considered a reliable predictor" baird2012 C36. The glass-stability parameters measure crystallization of the reheated liquid above Tg, where nucleation and growth are much faster than in the glass, and their inverse relation to Rcrit was assumed from inorganic glasses rather than shown for organic molecules baird2010 C43, C26. Celecoxib's inconsistent parameters are "believed to be" due to its high Tg and crystallization temperature relative to Tm baird2010 C29. For fragility the authors note that organic liquids are held together by non-covalent forces much weaker than covalent networks, so non-Arrhenius (fragile) behaviour is general among them, and on a log scale the large absolute viscosity differences between compounds shrink baird2012 C40, C42. Hence Angell plots "give minimal information" on crystallization tendency, and kinetic fragility "may not accurately assess" glass-forming ability; crystallization also depends on thermodynamic factors baird2012 C43, C44. The authors summarize that none of the inorganic-glass parameters correlated strongly with crystallization behaviour, "suggesting these parameters may not be ideal" for organic molecules baird2010 C38. A 2010 library paper on felodipine–PVP dispersions notes from the literature that crystallization tendency cannot be predicted from relaxation times and Tg alone marsac2010 C29. The formulas are on the reduced glass transition temperature and Vogel-Tammann-Fulcher equation pages.

Conditions and limits

No statistical test is named for the Trg comparisons, and 16 class I Tg values are estimates from molecular weight baird2010 R14, X17. Tred and the glass-stability parameters cannot be reproduced from the printed temperatures, and chlorpropamide's KSP and K2 are printed identical baird2010 X8, X1. The fragility category depends on whether m or D is used: aceclofenac, tolazamide and nifedipine are strong by m and D^a but fragile by D^b baird2010 X14. Class I-A was not fitted with VTF, and the fitted T0 values do not satisfy the paper's own relation between T0, D and Tg baird2012 X7, X5. All data are for neat melts, without polymer baird2010 A1; baird2012 A1.

Counterexamples and conflicts

The class means of Tg/Tm do follow the expected order, class I lowest and class III highest baird2010 C37. For crystal growth rates, slow growers had a small Tm − Tg trasi2014a R8 (synthesis: a small Tm − Tg means a high Tg/Tm, so the reduced-Tg idea keeps some value as a class-level trend for growth while failing to rank molecules by DSC class; see F5).

Transfer check

Do not screen candidates by Tg/Tm, glass-stability parameters or fragility alone baird2010 C38; baird2012 C36, C43. For a new drug, the heat–cool–heat test, a seeded growth rate and the melt viscosity at Tm with its rise over the first 40 °C of undercooling carried more information in these data sets baird2012 C45; baird2010 C64. If Tg/Tm is reported, measure Tg rather than estimating it from molecular weight baird2010 X17.

History

Trg goes back to Kauzmann and Turnbull, with a typical value of 0.66 proposed baird2010 R29, C1. Baird in 2010 tested Trg, the glass-stability parameters and calorimetric fragility on 51 molecules and found none of them reliable baird2010 C38. Baird in 2012 confirmed the fragility result with viscosity-based fits and showed that Tg/Tm misses differences in the rise of viscosity baird2012 C43, C36. Nguyen in 2023 still reported Tg/Tm alongside the class for two antitubercular drugs nguyen2023a C59.

F13

Topic 2Screening tests built on them5 of 6

Small, rigid molecules with large heats of fusion tend to crystallize fast, but no single property decides.

Two statistical analyses of seven molecular properties for 51 and 40 drugs, run without class labels, placed the fast crystallizers on average among smaller, more rigid molecules with larger heats of fusion, and the slow ones among larger, more flexible molecules. The separation was partial, with outliers in every class baird2010 R20, R22, C40, R21, C63; vaneerdenbrugh2010a R18, R20, C36, R19, C45.

moderateModerate: the analyses were cross-validated and built without class labels, but they describe class averages with outliers, and Tg was estimated for 16 molecules baird2010 K6, X17; vaneerdenbrugh2010a K5, X9.

class I: fast crystallizersclass III: glass formersmolecular weight−0.63+0.79rotatable bonds−0.54+0.66heat of fusion−0.63+0.45entropy of fusion−0.59+0.41−0.8−0.6−0.4−0.200.20.40.60.8class mean, in standard deviations from the overall mean

51 molecules classified from undercooled melts; fast crystallizers are smaller, more rigid and have larger heats and entropies of fusion. baird2010 R22

Class I (fast crystallizers)
benzocainelidocainehaloperidol
Close relative in another class
procainedroperidol

Procaine is class III and droperidol class II. baird2010 R10, C14

For your drug

  1. Use molecular size, flexibility and heat of fusion only as a prior: a small, rigid molecule with a high heat of fusion starts with a higher risk, a large flexible one with a lower risk.
  2. Measure the class itself, by the DSC cycle or the spin-coat test (F9, F14). If you use a PCA model, include the drug's measured Tg and check that the drug lies within the descriptor range of the 51 molecules.

baird2010 C40, C64; vaneerdenbrugh2010a C36, C47, C46

How it works and key numbers
  1. Flexible molecules exist in the melt as energetically similar conformers (shapes), and more rotatable bonds lower the chance that a molecule is in the orientation needed to join a nucleus or crystal baird2010 C59.
  2. Higher heat and entropy of fusion give a more negative ΔGv and so, by classical nucleation theory, a lower barrier (see F6); a higher melting point means more energy to disrupt the lattice baird2010 C60, C61, C62; vaneerdenbrugh2010a C38, C39.
  3. Because several properties were needed, the authors conclude that multiple properties "appear to play a role" and that no single descriptor can predict crystallization baird2010 C42, C63; vaneerdenbrugh2010a C45.
  4. Conformer populations, interfacial energies and nucleation rates were not measured baird2010 C59, C61.
  5. Exceptions exist. Nilutamide, small with one rotatable bond, "would be expected" to crystallize readily but is a slow crystallizer; the authors think the issue is which conformer the crystal needs. Haloperidol (375.9 g/mol) is class I-A, acetaminophen (151.2 g/mol) class II trasi2014b M1, C36, C5, C51; baird2013 X11.
−0.63 vs +0.79 SDmolecular weight of class I against class III, in standard deviations from the overall meanbaird2010 R22
two of threevalidation compounds placed in the right class by the per-class modelsvaneerdenbrugh2010a R22, C44
1 rotatable bondnilutamide, a small molecule that is nevertheless a slow crystallizertrasi2014b M1, C36

Connections to other chapters

Evidence: data, conditions and papers

Finding F13. Small, rigid molecules with large heats of fusion tend to crystallize fast, but no single descriptor decides

In short. Two principal component analyses of seven descriptors (melting point, Tg, ΔHfus, ΔSfus, ΔGv, molecular weight, rotatable bonds), built without class labels, placed rapid crystallizers on average among smaller, more rigid molecules with larger heats and entropies of fusion, and slow crystallizers among larger, more flexible ones; class II sat between baird2010 R20, R22, C40; vaneerdenbrugh2010a R18, R20, C36. Separation was partial, outliers occurred in every class, and no single descriptor sufficed baird2010 R21, C63; vaneerdenbrugh2010a R19, C45. Per-class models placed two of three validation compounds correctly, and close structural relatives fell in different classes vaneerdenbrugh2010a R22, C44, X8; baird2010 R10, C14.

Type. observation in N systems (51 and 40 compounds; 31 drugs for growth) Strength. cross-validated PCA built without class labels, but class averages with outliers and partly estimated Tg values (baird2010 K6, vaneerdenbrugh2010a K5: moderate).

How it was shown

Baird and co-workers (2010) ran a PCA on all 51 molecules with seven descriptors (ΔHfus, ΔSfus and ΔGv per unit volume, melting onset, Tg, molecular weight and rotatable bonds); no class information was given to the model baird2010 P8. Van Eerdenbrugh and co-workers (2010) repeated it for 40 drugs after spin coating, built one model per class and projected three compounds not used in the models vaneerdenbrugh2010a P11, P12.

What the data show

The melt model explained 97.8% of the descriptor variance with three components and predicted 91.1% in cross-validation, with outliers in every class baird2010 R20, C66, R21. PABA (class I) and clarithromycin (class III) were "well predicted" vaneerdenbrugh2010a R22, C44.

papersystemresultids
baird201051 molecules, melt classesclass I MW ≈−0.63 and rotatable bonds ≈−0.54 SD; class III ≈+0.79 and ≈+0.66baird2010 R20, R22, C40
vaneerdenbrugh2010a40 drugs, solvent classesMW ≈−0.68 / +0.70, rotatable bonds ≈−0.42 / +0.57, Tg ≈−0.34 / +0.46 for class I / IIIvaneerdenbrugh2010a R18, R20, C40
vaneerdenbrugh2010athree validation compoundsPABA (class I) and clarithromycin (class III) placed correctly; piroxicam (class II) fits both modelsvaneerdenbrugh2010a R22, C44, X8
baird2010structural relativesbenzocaine and lidocaine class I against procaine class III; haloperidol class I against droperidol class IIbaird2010 R10, C14
trasi2014a31 drugs, growth groupsfast growers lower MW (≈−0.28 SD), fewer rotatable bonds (≈−0.35), higher Tm (≈+0.55)trasi2014a R8, C25
Fig. 3 of baird2010: PCA scores of 51 molecules coloured by crystallization class
Fig. 3 · baird2010PCA scores of 51 molecules coloured by crystallization class

Class I (red) lies mostly left and low, class III (green) up and right and class II (blue) between; outliers of every class sit inside the other clusters baird2010 R21.

Why

The authors' explanation for size and flexibility is conformational: more rotatable bonds lower the chance that a molecule is in the orientation needed to join a nucleus or crystal baird2010 C59. Higher ΔHfus and ΔSfus give a more negative ΔGv and so, by classical nucleation theory, a lower barrier baird2010 C60, C61; vaneerdenbrugh2010a C38, C39. Conformer populations, interfacial energies and nucleation rates were not measured baird2010 C59, C61.

Limits and transfer

The trends are class averages with wide spread, and Tg was estimated for 16 molecules in both PCAs baird2010 R21, X17; vaneerdenbrugh2010a X9. The prediction was tested on three compounds only vaneerdenbrugh2010a X8. Use size, flexibility and ΔHfus only as a prior for a new drug; the class itself has to be measured, by the DSC cycle or the spin-coat test baird2010 C40, C64; vaneerdenbrugh2010a C36, C47.

How it was shown

Baird and co-workers (2010) ran a PCA (SIMCA-P+) on all 51 molecules of their DSC screen with seven descriptors: melting onset, Tg midpoint, ΔHfus, ΔSfus and ΔGv (Hoffman at 293.15 K), the last three per unit volume via the crystal density, molecular weight and number of rotatable bonds baird2010 P8. Data were centred and scaled, components were chosen by cross-validation, and no class information was given to the model baird2010 P8. For 16 class I molecules that crystallized too fast to show a Tg, Tg was estimated from a regression of Tg/Tm on molecular weight baird2010 P8, E27, X17. Van Eerdenbrugh and co-workers (2010) repeated the analysis for the 40 drugs of their spin-coating screen with the same descriptors, then built one PCA model per class and projected three compounds not used in the models (PABA, piroxicam, clarithromycin) onto them in a Coomans' plot vaneerdenbrugh2010a P11, P12. Class membership was compared with the descriptors as each class's mean deviation from the overall mean in standard deviations baird2010 R22; vaneerdenbrugh2010a R20. Trasi and co-workers (2014) did the same for three groups of drugs sorted by crystal growth rate, over 11 properties trasi2014a P6, E5, R8.

What the data show

The 2010 melt model explained 97.8% of the descriptor variance with three components (56.3, 30.4 and 11.0%) and predicted 91.1% in cross-validation baird2010 R20, C66. On the first two components, class means lay at about (−1.1, −0.3) for class I, (0.0, 0.0) for class II and (1.4, 0.4) for class III, with outliers in every class (most notably lidocaine and tolbutamide in class I) baird2010 R21, C39. Class I had lower molecular weight (about −0.63 SD) and fewer rotatable bonds (−0.54), higher ΔHfus (+0.45), ΔSfus (+0.41) and melting onset (+0.25), and a more negative ΔGv (−0.40); class III showed the opposite signs (MW +0.79, rotatable bonds +0.66, ΔHfus −0.63, ΔSfus −0.59, melting onset −0.29, ΔGv +0.57) baird2010 R22. Tg barely differed between classes (all within ±0.08 SD) baird2010 R22.e, C41. The solvent model explained 97.6% (89.8% cross-validated), with class means at about (−0.9, −0.6), (0.4, −0.2) and (0.7, 0.7) vaneerdenbrugh2010a R18, R19. Its class contributions ran MW −0.68 / +0.70, rotatable bonds −0.42 / +0.57, ΔHfus +0.41 / −0.26 and ΔSfus +0.46 / −0.33 for class I / III, and here Tg did separate the classes (−0.34 / +0.46) vaneerdenbrugh2010a R20, C40. The per-class models explained 97.9, 99.1 and 96.1% for classes I, II and III; the class III model could not discriminate and was not used vaneerdenbrugh2010a R21, C42. PABA (class I) and clarithromycin (class III) were "well predicted", while piroxicam (class II) fitted both models vaneerdenbrugh2010a R22, C44. Among growth groups, fast growers had lower molecular weight (about −0.28 SD), fewer rotatable bonds (−0.35) and higher Tm (+0.55) trasi2014a R8, C25. Close relatives split: benzocaine and lidocaine were class I but procaine class III, haloperidol class I but droperidol class II baird2010 R10, C14. Haloperidol (375.9 g/mol) is class I-A and acetaminophen (151.2 g/mol) class II, against class means of 217 g/mol for I-A and 324 g/mol for II in the 2013 set baird2013 X11. Nilutamide, small with one rotatable bond, "would be expected" to crystallize readily but is a slowly crystallizing good glass former trasi2014b M1, C36, C5.

papersystemmethod and conditionsresultids
baird201051 molecules, melt classesPCA, 3 components, 97.8% explained, 91.1% cross-validatedclass I MW ≈−0.63 and rotatable bonds ≈−0.54 SD; class III ≈+0.79 and ≈+0.66R20, R22, C40, C42
vaneerdenbrugh2010a40 drugs, solvent classesPCA, 97.6% explained, 89.8% cross-validatedMW ≈−0.68 / +0.70, rotatable bonds ≈−0.42 / +0.57, Tg ≈−0.34 / +0.46 for class I / IIIR18, R20, C36, C40
vaneerdenbrugh2010athree validation compoundsper-class PCA models, Coomans' plotPABA (class I) and clarithromycin (class III) placed correctly; piroxicam (class II) fits both modelsR22, C44, X8
baird2010structural relativesDSC screenbenzocaine and lidocaine class I against procaine class III; haloperidol class I against droperidol class IIR10, C14
baird201325 compoundsconclusionclass I-A "tend to be" low-MW, simple molecules; exceptions haloperidol (375.9 g/mol, I-A) and acetaminophen (151.2 g/mol, II)C39, X11
trasi2014a31 drugs, growth groupsstandardized group deviationsfast growers lower MW (≈−0.28 SD), fewer rotatable bonds (≈−0.35), higher Tm (≈+0.55)R8, C25
trasi2014bnilutamide—a small molecule with one rotatable bond "would be expected" to crystallize readily, yet it is a slowly crystallizing good glass formerM1, C36, C5
nguyen2023apretomanid—high crystallization tendency attributed to low Tg, low MW and a relatively linear structureC61
ilevbare2013critonavir against celecoxib and efavirenzsupersaturated bufferslower nucleation of ritonavir attributed to higher MW and greater conformational flexibilityC25
hsieh2014four weak basespH-induced precipitationconformational-freedom argument for crystallization does not follow the printed rotatable-bond countsC33, X12

Figure

Fig. 3 of baird2010: PCA scores of 51 molecules coloured by crystallization class
Fig. 3 · baird2010PCA scores of 51 molecules coloured by crystallization class

Class I (red) lies mostly left and low, class III (green) up and right and class II (blue) between; triangles are class means, and outliers of every class sit inside the other clusters baird2010 R21.

Fig. 4 of vaneerdenbrugh2010a: class contributions of the seven descriptors after spin coating
Fig. 4 · vaneerdenbrugh2010aclass contributions of the seven descriptors after spin coating

Black (class I) bars point down for molecular weight, rotatable bonds and Tg and up for ΔHfus and ΔSfus; white (class III) bars point the other way vaneerdenbrugh2010a R20.

Why

The authors' explanation for size and flexibility is conformational: flexible molecules exist in the melt as energetically similar conformers, and more rotatable bonds lower the chance that a molecule is in the orientation needed to join a nucleus or crystal baird2010 C59. Van Eerdenbrugh and co-workers call this "intuitive" vaneerdenbrugh2010a C37. The thermal descriptors are read through the driving force: higher ΔHfus and ΔSfus give a more negative ΔGv and so, by classical nucleation theory, a lower barrier (see F6) baird2010 C60, C61; vaneerdenbrugh2010a C38, C39. A higher melting point means more energy to disrupt the lattice baird2010 C62. Conformer populations, interfacial energies and nucleation rates were not measured baird2010 C59, C61. Tg separated the solvent classes but not the melt classes because a week of storage at room temperature favours compounds whose Tg is above the storage temperature, whereas the melt test takes every sample above Tg ("can be readily understood") vaneerdenbrugh2010a C41; baird2010 C41. Because several descriptors were needed, the authors conclude that multiple properties "appear to play a role" and that no single descriptor can predict crystallization baird2010 C42, C63; vaneerdenbrugh2010a C45. Other library papers use the same reasoning for single drugs: pretomanid's high tendency is "attributed" to its low Tg, low molecular weight and linear structure, and ritonavir's slow solution nucleation "most likely" to its size and flexibility nguyen2023a C61; ilevbare2013c C25. For nilutamide the authors propose that the issue is which conformer the crystal needs, not the number of rotatable bonds (see F7) trasi2014b C51. (synthesis: the descriptor trends describe the average of a class, and both PCAs share estimated Tg values for the fastest crystallizers, so the trends are a prior for a new drug rather than a model of it baird2010 X17; vaneerdenbrugh2010a X9.)

Conditions and limits

Class-average trends with wide spread baird2010 R21; vaneerdenbrugh2010a R19. Tg was estimated for 16 molecules in both PCAs baird2010 X17; vaneerdenbrugh2010a X9. Per-compound descriptor values are not printed in the solvent study, and its class bars are plot readings vaneerdenbrugh2010a R20. The prediction was tested on three compounds only vaneerdenbrugh2010a X8. The descriptor set holds no measure of hydrogen bonding or crystal packing beyond ΔHfus and Tm baird2010 P8.

Counterexamples and conflicts

Nilutamide trasi2014b C36; haloperidol and acetaminophen baird2013 X11; structural relatives in different classes baird2010 R10. For four weak bases a reduced-conformational-freedom argument did not follow the printed rotatable-bond counts hsieh2014 X12. Polyphenols with many hydroxyl groups (naringenin, quercetin) crystallized readily, which the authors attribute to an extensive hydrogen-bonded network rather than to size wegiel2013a C26.

Transfer check

Use molecular size, flexibility and ΔHfus only as a prior for a new drug: a small, rigid molecule with a high heat of fusion starts with a higher risk, a large flexible one with a lower one baird2010 C40; vaneerdenbrugh2010a C36. The class itself has to be measured, by the DSC cycle or the spin-coat test (see F9, F14) baird2010 C64; vaneerdenbrugh2010a C47. If a PCA model is used, include the drug's measured Tg and check that the drug falls within the descriptor range of the 51 molecules vaneerdenbrugh2010a C46.

History

Both PCAs date from 2010 baird2010 C40; vaneerdenbrugh2010a C36. Baird in 2013 restated the low-MW tendency baird2013 C39. Trasi in 2014 found similar descriptors for fast growth and a small, rigid exception in nilutamide trasi2014a C25; trasi2014b C36. Later papers used size and flexibility to explain slow solution nucleation of ritonavir (2013) and fast crystallization of pretomanid (2023) ilevbare2013c C25; nguyen2023a C61.

F14

Topic 2Screening tests built on them6 of 6

Fast drying gives the same class whatever the solvent or route, but slow drying crystallizes most slow crystallizers.

Spin-coating a drug into films about 1 µm thick and storing them dry for a week sorted 40 drugs into rapid, intermediate and slow crystallizers. Changing the solvent changed the class of only 4 of 33 drugs vaneerdenbrugh2010a R9, R11. When the solvent was dried off slowly in a vial instead, 22 of 31 slow-crystallizer combinations were fully crystalline at once vaneerdenbrugh2010a R13.

moderateModerate overall: the class counts, the small solvent effect and the drying-rate contrast are direct and clear, but the match with the melt rests on cited classes with different criteria, and crystallinity was scored by eye vaneerdenbrugh2010a K1, K2, K3, K4, P8.

Spin-coat against melt class, % of 93 combinationssame class68%one class apart30%opposite classesonly atenolol2%

No combination was class I by evaporation and class III from the melt. vaneerdenbrugh2010a R16, R17

Slow-crystallizer combinations dried slowlycombinations tested31fully crystalline at onceincl. celecoxib, felodipine, indomethacin22

Class III combinations dried slowly in a vial; the same combinations stayed mostly amorphous for a week after spin coating. vaneerdenbrugh2010a R12, R13

For your drug

  1. A spin-coat screen of the neat drug ranks it for fast-drying processes such as spray drying. For slow solvent removal (rotary evaporation, film casting, slow tray drying), test the drug at the process drying rate.
  2. If the drug falls in the intermediate class, or its class changes with solvent, test it in the actual process solvent. Check Tg against the storage temperature, since a Tg above it went with the slow class after evaporation.

vaneerdenbrugh2010a C12, C32, R13, R11, R14, R20, C41

How it works and key numbers
  1. Spin coating dries a thin film in seconds, which would suppress any solvent-specific effect on nucleation and growth ("can be explained") vaneerdenbrugh2010a C25, C11.
  2. The authors read class I as extremely fast nucleation and growth and class III as slow nucleation and/or growth; none of these rates was measured vaneerdenbrugh2010a C18, C22.
  3. From the agreement with the melt classes they conclude ("suggests") that under rapid solidification the drug's own properties dominate vaneerdenbrugh2010a C33.
  4. Slow evaporation keeps the solution supersaturated longer before its viscosity rises, so crystallization becomes "an obvious result", and the solvent then matters more vaneerdenbrugh2010a C27, C28.
  5. Both routes are fast, so their agreement shows that the drug ranks alike under fast solidification, not that the process is irrelevant (the extractor's reading) vaneerdenbrugh2010a X7, R13.
  6. Later work met the same limit: rotary evaporation gave no amorphous naringenin, and delamanid dispersions above about 10% drug loading held crystals, put down to an evaporation rate "not fast enough" wegiel2013a R7; duong2022a C44, C45.
4 of 33drugs whose class changed with the solventvaneerdenbrugh2010a R11
68%of 93 drug–solvent combinations in the same class by spin coating and by meltvaneerdenbrugh2010a R16, R17
22 of 31slow-crystallizer combinations fully crystalline at once when dried slowlyvaneerdenbrugh2010a R13

Connections to other chapters

2 more connections
Evidence: data, conditions and papers

Finding F14. Under fast solidification the route and the solvent hardly change the class; slow solvent removal crystallizes most slow crystallizers

In short. Spin-coated into films about 1 µm thick and stored dry for a week, 40 drugs sorted into rapid, intermediate and slow crystallizers, and changing among three solvents changed the class of only 4 of 33 drugs vaneerdenbrugh2010a R9, R11. The spin-coating class equalled the melt class of the 2010 DSC screen in 68% of drug–solvent combinations and was opposite only for atenolol vaneerdenbrugh2010a R16, R17. Drying rate mattered more than solvent or route: dried slowly in a vial, 22 of 31 slow-crystallizer combinations were fully crystalline at once, and later library papers could not make neat amorphous naringenin, curcumin or high-load delamanid dispersions by rotary evaporation vaneerdenbrugh2010a R13; wegiel2013a R7; wegiel2014 C9; duong2022a C45.

Type. observation in N systems (40 drugs, 93 drug–solvent combinations; three later drugs) Strength. the class counts, the small solvent effect and the drying-rate contrast are direct and clear (vaneerdenbrugh2010a K1, K2, K3: strong), but the match with the melt rests on cited melt classes with different criteria (K4: moderate) and on visual scoring (section 0.8: moderate).

How it was shown

Van Eerdenbrugh, Baird and Taylor (2010) measured the solubility of the 51 melt-screen compounds in dichloromethane, ethanol and a 1:1 mixture and kept 93 of 153 combinations of 40 drugs vaneerdenbrugh2010a P5, P6, R5, R6, R7. 200 µL of solution was spun for 20 s at 8000 rpm, giving films about 1 µm thick, scored by polarized light over 7 days of dry storage vaneerdenbrugh2010a P2, P8. For class III combinations, 400 µL was also left to evaporate slowly vaneerdenbrugh2010a P7.

What the data show

Only 4 of 33 drugs tested in more than one solvent changed class vaneerdenbrugh2010a R11. Against the melt classes, 68% of combinations were identical and 2% opposite, all atenolol vaneerdenbrugh2010a R16, R17. Slow evaporation made 22 of the 31 class III combinations fully crystalline at the first look vaneerdenbrugh2010a R12, R13.

papersystemmethod and conditionsresultids
vaneerdenbrugh2010a40 drugs, 93 combinations0.26 M in DCM, MIX or EtOH; spin coating; PLM over 7 days dry39, 23 and 31 in classes I, II and III; 4 of 33 drugs changed class with solventvaneerdenbrugh2010a P1, P2, R9, R11
vaneerdenbrugh2010aagainst the 2010 melt classescross-tabulation of 93 combinationsidentical 68%, one class apart 30%, opposite 2% (atenolol); per drug 25 of 40 agree in every solventvaneerdenbrugh2010a R16, R17, X6
vaneerdenbrugh2010a31 class III combinationsslow evaporation of 400 µL in a vial22 of 31 fully crystalline at day 0; clotrimazole, ketoprofen, miconazole, ritonavir solvent dependentvaneerdenbrugh2010a R12, R13
wegiel2013anaringenin, quercetinrotary evaporation; spin coatingno amorphous material by rotary evaporation; amorphous naringenin, not quercetin, by spin coatingwegiel2013a R7, C19
duong2022adelamanid dispersionsrotary evaporationresidual crystallinity above about 10–15% drug loadingduong2022a R8, C45
Table 4 of vaneerdenbrugh2010a: spin-coating class against melt class, % of 93 combinations
Table 4 · vaneerdenbrugh2010aspin-coating class against melt class, % of 93 combinations

The diagonal (35, 8 and 25%) is the 68% agreement; the class I evaporation, class III melt cell is empty and the opposite cell holds only atenolol vaneerdenbrugh2010a R16, R17.

Why

The authors explain the small solvent effect by the very short drying time of spin coating, which would suppress any solvent-specific effect ("can be explained") vaneerdenbrugh2010a C25. Slow evaporation keeps the solution supersaturated longer, so crystallization becomes "an obvious result" vaneerdenbrugh2010a C27, C28. (synthesis: the drying rate can override the class vaneerdenbrugh2010a X7, R13.)

Limits and transfer

Crystallinity was scored by eye, and the melt classes are cited from the 2010 DSC study with different criteria and time scales vaneerdenbrugh2010a P8, R10, P10, C30. Slow evaporation was run on class III combinations only vaneerdenbrugh2010a P7. For slow solvent removal, test the drug at the process drying rate vaneerdenbrugh2010a R13.

How it was shown

Van Eerdenbrugh, Baird and Taylor (2010) asked how readily drugs crystallize after rapid solvent evaporation, whether the solvent or the route to the amorphous state changes that, and which compound properties go with it vaneerdenbrugh2010a A1, A2, A4. They took the 51 compounds of the melt screen, measured their solubility in dichloromethane (DCM), ethanol (EtOH) and a 1:1 (w/w) mixture (MIX), and removed combinations that degraded in solution (griseofulvin, nifedipine, procaine, tolfenamic acid) or sublimed under vacuum (antipyrin, ibuprofen), keeping 93 of 153 combinations of 40 drugs vaneerdenbrugh2010a P5, P6, R5, R6, R7. A standard concentration of 0.26 M was chosen with felodipine, because below it the films crystallized only in patches at 94% RH; six combinations were run at 0.20 M vaneerdenbrugh2010a P3, P4, R3, P1. 200 µL of solution was spun for 20 s at 8000 rpm on an 18 × 18 mm cover slip, in triplicate, giving films about 1 µm thick, which the authors liken to spray drying ("some similarity") vaneerdenbrugh2010a P2, C12. Films were scored by polarized light within 30 min (day 0), after a night under vacuum (day 1) and after 2, 4 and 6 more days over P2O5 at room temperature (days 3, 5, 7), on a five-step scale from AAAA (no birefringence) to CCCC (fully crystalline) vaneerdenbrugh2010a P8. Class I meant at least AACC at day 0, class II at least AACC by day 7, class III AAAA or AAAC at day 7 vaneerdenbrugh2010a P9. For every class III combination, 400 µL of the same solution was also left to evaporate at ambient conditions in a glass vessel 12 mm wide and 6 mm high, in triplicate vaneerdenbrugh2010a P7. The spin-coating classes were compared with the melt classes of the 2010 DSC screen, which used cooling at 20 °C/min and reheating at 10 °C/min vaneerdenbrugh2010a P10; baird2010 P1.

What the data show

The 93 combinations split into 39 class I, 23 class II and 31 class III; by compound, 14 were class I, 9 class II and 13 class III in every solvent tested, and clofoctol, flurbiprofen (I or II), cinnarizine and loratadine (II or III) changed class with the solvent vaneerdenbrugh2010a R8, R9. Of the 33 drugs tested in more than one solvent, 10 differed in category at one or more time points, but only those 4 changed class vaneerdenbrugh2010a R11. About half of the class II drugs showed a little crystallinity right after spin coating, and no class III drug did vaneerdenbrugh2010a R10. Against the melt classes, 35% of the 93 combinations were class I by both routes, 8% class II by both and 25% class III by both, 68% in all; 30% were one class apart and 2% opposite, the 2% being atenolol in MIX and EtOH (spin-coat class III, melt class I) vaneerdenbrugh2010a R16, R17. No combination was class I by evaporation and class III from the melt vaneerdenbrugh2010a R16. Counted per drug, 13 of 14 spin-coat class I drugs were also melt class I and 10 of 13 spin-coat class III drugs melt class III, but only 2 of 9 spin-coat class II drugs were melt class II; 25 of 40 drugs agree in every solvent vaneerdenbrugh2010a R14, X6. Slow evaporation reversed the picture for slow crystallizers: 22 of the 31 class III combinations were CCCC at the first look, including celecoxib, felodipine and indomethacin in every solvent tested vaneerdenbrugh2010a R12, R13. Only clotrimazole, ketoprofen, miconazole and ritonavir were not fully crystalline at once, and their outcome then depended on the solvent: miconazole was CCCC from DCM but AAAA for 7 days from MIX, and ritonavir CCCC from DCM but AAAA to AACC from MIX vaneerdenbrugh2010a R12, R13. Later work met the same drying-rate limit: rotary evaporation gave no amorphous naringenin or quercetin, while spin coating gave amorphous naringenin; curcumin could be made amorphous by spin coating but not in larger amounts by rotary evaporation wegiel2013a R7; wegiel2014 C9. For delamanid, a class I drug, rotary-evaporated dispersions above about 10% drug loading held crystals, which the authors put down to an evaporation rate "not fast enough" duong2022a C44, C45.

papersystemmethod and conditionsresultids
vaneerdenbrugh2010a40 drugs, 93 combinations0.26 M in DCM, MIX or EtOH; spin coating 20 s at 8000 rpm; PLM over 7 days dry39 / 23 / 31 in classes I / II / III; 4 of 33 drugs changed class with solventP1, P2, R9, R11
vaneerdenbrugh2010aagainst the 2010 melt classescross-tabulation, % of 93 combinationsidentical 68%, one class apart 30%, opposite 2% (atenolol); per drug 25 of 40 agree in every solventR16, R17, X6
vaneerdenbrugh2010a31 class III combinationsslow evaporation of 400 µL in a vial22 of 31 CCCC at day 0; clotrimazole, ketoprofen, miconazole, ritonavir solvent dependentR12, R13, C26, C28
baird2010the same compounds, melt routeDSC, cool 20 °C/min, reheat 10 °C/min23 / 11 / 17 of 51 molecules in classes I / II / IIIP1, R1
vaneerdenbrugh2010bdrugs with polymersrotary evaporation against spin coating (argument)rotary evaporation is much slower and at 35 °C, lengthening the time for nucleation and growth in solutionC39
wegiel2013anaringenin, quercetinrotary evaporation; spin coatingno amorphous material by rotary evaporation; amorphous naringenin, not quercetin, by spin coatingR7, C19
wegiel2014curcuminspin coating against rotary evaporationamorphous by spin coating, not in larger amounts by rotary evaporationC9
duong2022adelamanid dispersionsrotary evaporation, HPMCAS and HPMCPresidual crystallinity above about 10–15% drug loading; spray drying "It is likely" to avoid itR8, C45, C46

Figure

Table 3 of vaneerdenbrugh2010a: slow crystallizers dried slowly, crystallinity from day 0 to day 7
Table 3 · vaneerdenbrugh2010aslow crystallizers dried slowly, crystallinity from day 0 to day 7

Most rows read CCCC from day 0 onward, although every one of these combinations stayed mostly amorphous for a week after spin coating; the four exceptions vary with the solvent vaneerdenbrugh2010a R12, R13.

Table 4 of vaneerdenbrugh2010a: spin-coating class against melt class, % of 93 combinations
Table 4 · vaneerdenbrugh2010aspin-coating class against melt class, % of 93 combinations

The diagonal (35, 8 and 25%) is the 68% agreement; the class I evaporation, class III melt cell is empty and the opposite cell holds only atenolol vaneerdenbrugh2010a R16, R17.

Why

The authors explain the small solvent effect by the very short drying time of spin coating, which would suppress any solvent-specific effect on nucleation or growth ("can be explained") vaneerdenbrugh2010a C25. The thin film and the fast process "should minimize" nucleation and growth in the supersaturated solution vaneerdenbrugh2010a C11. Slow evaporation keeps the solution supersaturated longer before its viscosity rises, so crystallization becomes "an obvious result", and the solvent then matters more ("implies") vaneerdenbrugh2010a C27, C28. They read the classes kinetically: class I crystallized at once, "suggesting" extremely fast nucleation and growth; class II crystals come from nuclei formed during spin coating, from nucleation in the film, or both ("may be due", "probably important"); class III "implying" slow nucleation and/or growth vaneerdenbrugh2010a C18, C20, C22. None of these rates was measured, and the duration of supersaturation was not measured either vaneerdenbrugh2010a C18, C22, C25. From the agreement with the melt classes they conclude ("suggests") that under rapid solidification crystallization tendency is dominated by the physicochemical properties of the drug vaneerdenbrugh2010a C33. Part of the difference between the routes is built into the definitions: the melt test is a short temperature excursion above Tg, while the solvent test is a week of storage, which favours drugs whose Tg lies above room temperature ("can be readily understood") vaneerdenbrugh2010a C30, C41. Duong and co-workers later state the same principle for dispersions: the evaporation rate decides whether a drug is trapped before nucleation and growth ("Clearly") duong2022a C45. (synthesis: both 2010 routes are fast, so their agreement shows that the drug ranks alike under fast solidification, not that the process is irrelevant; the slow-evaporation result shows that the drying rate can override the class vaneerdenbrugh2010a X7, R13.)

Conditions and limits

Neat drugs, three solvents of different polarity, mostly 0.26 M, films about 1 µm thick, room temperature and dry storage for 7 days vaneerdenbrugh2010a P1, P2, P8. Crystallinity was scored by eye in five steps, and the per-sample data are in a supporting information file that was not available vaneerdenbrugh2010a P8, R10. The melt classes are cited from the 2010 DSC study and defined by different criteria and time scales vaneerdenbrugh2010a P10, C30. Slow evaporation was run on the class III combinations only vaneerdenbrugh2010a P7. Polymers, humidity, higher temperatures and storage beyond 7 days were not part of the study vaneerdenbrugh2010a A1, A5, P8. Eleven of the 51 compounds were dropped, five of them without a stated reason vaneerdenbrugh2010a X3.

Counterexamples and conflicts

Atenolol is spin-coat class III but melt class I vaneerdenbrugh2010a R17. The intermediate class agrees poorly between routes: of 9 spin-coat class II drugs, 3 were melt class I and 4 melt class III vaneerdenbrugh2010a R14. Per drug the agreement is 25 of 40, lower than the 68% per combination vaneerdenbrugh2010a X6. The statement that the compound rather than the processing decides holds only across the two rapid routes, not across drying rates vaneerdenbrugh2010a X7.

Transfer check

A spin-coat screen of the neat drug ranks it for fast-drying processes such as spray drying; the authors expect a drug that crystallizes readily during solvent evaporation also to crystallize easily from a melt vaneerdenbrugh2010a C12, C32. For slow solvent removal (rotary evaporation, film casting, slow tray drying), test the drug at the process drying rate, because 22 of 31 slow crystallizers became fully crystalline when dried slowly vaneerdenbrugh2010a R13. A drug in the intermediate class, or one whose class changes with solvent, should be tested in the actual process solvent vaneerdenbrugh2010a R11, R14. Check Tg against the storage temperature, since a Tg above it went with the slow class after evaporation vaneerdenbrugh2010a R20, C41.

History

Baird and co-workers in 2010 classified the 51 compounds from the melt baird2010 R1. Van Eerdenbrugh and co-workers in the same year established the solvent classes, their agreement with the melt classes and the drying-rate limit vaneerdenbrugh2010a C31, C26, and used the classes that year to choose drugs for a polymer screen vaneerdenbrugh2010b R11. Polyphenol studies in 2013 and 2014 met the drying-rate limit with rotary evaporation wegiel2013a R7; wegiel2014 C9. Duong in 2022 made the evaporation rate the explanation for residual crystallinity in delamanid dispersions duong2022a C45.

Topic 3 of 4 · 3 findings · F15–F17

Distinction between forming a glass and keeping it

F15

Topic 3Distinction between forming a glass and keeping it1 of 3

A drug that forms a glass easily may still crystallize quickly when the glass is reheated.

Ease of glass formation on cooling and resistance of the glass to crystallization are different properties. In a rapid calorimeter acetaminophen formed a glass below 1 °C/min yet still crystallized on reheating at 1910 °C/min baird2013 R9.e, R10, C24, C25. Among drugs that crystallized neither on cooling nor on reheating (class III), maximum growth rates ran from 216 to 0.02 µm/s, so for the fast growers glass formation reflects slow nucleation, not slow growth trasi2014a R1, C14, C15.

moderateModerate: the gap between cooling and reheating is large and direct, but the class III growth spread rests on values of unstated origin, and the low nucleation of the fast growers is inferred, not measured baird2013 K4; trasi2014a K2, X3, C15.

acetaminophen: glass below 1class I-B: glass at 25acetaminophen: still crystallizes at 1910gap of more than three orders of magnitude1101001000cooling or reheating rate, °C/min (log scale)

Cooling rate that gives a glass against reheating rate that still crystallizes it. baird2013 R9.e, R9, C24

Class III drugs (19): growth groups by maximum growth velocityfast growers7intermediate growers4slow growers8

Maximum velocities of class III run from 216 to 0.02 µm/s; the 12 class I and II drugs had 1.8–104 µm/s and none was a slow grower. trasi2014a R1, C14

For your drug

  1. Always reheat the glass and measure a seeded growth rate; a clean cooling trace is not evidence of stability.
  2. Treat a drug that forms a glass on slow cooling but crystallizes on reheating at 10–20 °C/min (class II), or that grows faster than about 10 µm/s at its maximum, as at risk once nuclei are present, for example after milling or incomplete melting. With a fast-scanning calorimeter, compare Rcrit,cool with Rcrit,heat.

baird2010 C47, R1; trasi2014a C16, C17; baird2013 C24

How it works and key numbers
  1. In the authors' reading, an easy glass former has nucleation and growth windows that overlap little: on cooling the melt passes the growth maximum while few nuclei exist, and forms nuclei only where growth is slow baird2013 C35, C36.
  2. On reheating, those nuclei and any formed on the way up are carried into the fast-growth range, where a rate that sufficed on cooling is too slow to stop growth baird2013 C36.
  3. For class III fast growers, ready glass formation can come from low nucleation, low growth, both, or poor overlap; the fast growers "clearly" form glasses because they nucleate poorly under the test conditions. Nucleation rates were not measured trasi2014a C13, C15.
  4. Nilutamide is read the same way: a low nucleation tendency, while its crystals grow fast ("it appears") trasi2014b C9.
  5. An extended classification separates the cases trasi2014a C17:
Easy glass, fast growth (class II)
nimesulidenilutamidecelecoxib
Slow nucleation and growth (class IV)
warfarinmiconazole

trasi2014a C17

below 1 vs above 1910 °C/minacetaminophen: cooling rate that gives a glass against reheating rate that still crystallizes itbaird2013 R9.e, C24
216 → 0.02 µm/sspread of maximum growth rates among class III drugstrasi2014a R1, C14
25 vs above 1500–2000 °C/minfour class I-B compounds: cooling that still gives a glass against reheating that avoids crystallizationbaird2013 R9, C24
Evidence: data, conditions and papers

Finding F15. Forming a glass easily does not make the glass resistant to crystallization

In short. Ease of glass formation on cooling and resistance of the glass to crystallization are different properties baird2010 C47; baird2013 C24. Class I-B and II glasses needed reheating rates far above the cooling rates that formed them: acetaminophen formed a glass below 1 °C/min yet still crystallized on reheating at 1910 °C/min, and flufenamic acid made glassy at 25 or 1025 °C/min crystallized on reheating at 1900 °C/min baird2013 R9.e, R10, C24, C25. Among drugs that crystallized neither on cooling nor on reheating (class III), maximum growth rates spanned the whole range from 216 to 0.02 µm/s, so their glass formation reflects slow nucleation rather than slow growth; nilutamide is one such case trasi2014a R1, C14, C15; trasi2014b C9.

Type. observation in N systems (25 molecules by rapid heat–cool DSC; 31 drugs by growth rate; nilutamide) Strength. the cooling–heating asymmetry is large and direct (baird2013 K4: strong), while the class III growth spread rests on MCVs of unstated origin (trasi2014a K2: moderate to strong; X3) and the low nucleation of fast growers is inferred (trasi2014a C15).

How it was shown

Baird and co-workers (2010) defined glass-forming ability as the ease of vitrifying on cooling and glass stability as resistance of the glass to crystallization baird2010 C4. In 2013 they measured Rcrit,cool and Rcrit,heat in a rapid heat/cool calorimeter for 4 class I-B and 10 class II compounds baird2013 P2, P4, R9. Trasi and co-workers (2014) measured seeded growth of 31 drugs and coloured each by its 2010 class trasi2014a P1, R1.

What the data show

For acetaminophen, Rcrit,heat exceeded Rcrit,cool by more than three orders of magnitude baird2013 R9, C22, R9.e. Among the 31 drugs, all 8 slow growers belonged to class III, as did 7 fast growers trasi2014a R1, C14.

papersystemresultids
baird20134 class I-B, 10 class II; rapid heat–cool DSCRcrit,cool 25 °C/min (I-B) or 5 °C/min and below (II), against Rcrit,heat above 1500–2000 °C/min (I-B) and 12.5 to above 1910 °C/min (II)baird2013 R9, C22, C24
baird2013flufenamic acid, cooled at 25 or 1025 °C/minglass transition on reheating; crystallization on reheating even at 1900 °C/minbaird2013 R10, C25
trasi2014a19 class III and 12 class I/II drugs, seeded melt growthclass III MCV 216 to 0.02 µm/s; class I/II 1.8–104 µm/strasi2014a R1, C14, C15
trasi2014bnilutamide, DSC and seeded growthglass on cooling without recrystallization on reheating; growth about 38–40 µm/s at 100 °Ctrasi2014b R2, R8
mosqueragiraldo2014celecoxib, hot-stage microscopysamples hard to nucleate; easily rendered amorphous from the meltmosqueragiraldo2014 R15, C7
Fig. 2 of baird2013: flufenamic acid forms a glass at 25 °C/min yet crystallizes on reheating at 1900 °C/min
Fig. 2 · baird2013flufenamic acid forms a glass at 25 °C/min yet crystallizes on reheating at 1900 °C/min

Panel (a) has no exotherm on cooling and a glass transition on reheating; panel (b), after cooling at 1025 °C/min, still shows an exotherm and a melting endotherm on reheating at 1900 °C/min baird2013 R10.

Why

In the authors' reading, a melt that forms a glass easily has nucleation and growth windows that overlap little: on cooling it passes the growth maximum while few nuclei exist baird2013 C35, C36. For fast-growing class III drugs, Trasi and co-workers argue that they "clearly" form glasses because they nucleate poorly under the test conditions; nucleation rates were not measured trasi2014a C13, C15.

Limits and transfer

The drugs were neat, with DSC scans lasting minutes baird2013 X15; trasi2014a P1. The 2014 growth rates carry no replicates, and reheating rates of 1900–2000 °C/min exceed the calibrated range of the calorimeter trasi2014a X17; baird2013 X10. Always reheat the glass and measure a seeded growth rate baird2010 C47; trasi2014a C16.

How it was shown

Baird and co-workers (2010) defined glass-forming ability as the ease of vitrifying on cooling and glass stability as resistance to crystallization of the glass, noting that "glass stability" as usually measured on reheating is "somewhat of a misnomer", since it measures the reheated undercooled liquid baird2010 C4. In their screen, class II glasses were reheated at 10 and 20 °C/min, and class I-B glasses made by liquid-N2 quenching were reheated in the Raman experiments baird2010 P4, R8, R15. In 2013 the same group measured, in a rapid heat/cool calorimeter, both the slowest cooling that avoids crystallization (Rcrit,cool) and the slowest reheating that avoids it (Rcrit,heat), for 4 class I-B and 10 class II compounds baird2013 P2, P4, R9. Trasi and co-workers (2014) measured seeded growth rates of 31 drugs from the undercooled melt and coloured each by its glass-forming class from the 2010 test, red for classes I and II and blue for class III trasi2014a P1, R1. Trasi and Taylor (2014) compared nilutamide's glass formation on cooling with the growth rate of its seeded crystals trasi2014b R2, R8.

What the data show

In the rapid calorimeter, four class I-B compounds formed glasses at 25 °C/min but needed above 1500–2000 °C/min on reheating baird2013 R9, C24. Class II compounds had Rcrit,cool of 5 °C/min or less but Rcrit,heat from 12.5 (bifonazole) to above 1910 °C/min (acetaminophen) baird2013 R9, C22. For acetaminophen the gap exceeds three orders of magnitude (below 1 against above 1910 °C/min), which the authors believe is the first report of such an asymmetry for organic systems ("to the best of the authors' knowledge"), similar ones being known in bulk metallic glasses baird2013 R9.e, C24, C26, R13. Flufenamic acid cooled at 25 °C/min gave no exotherm and a glass transition on reheating, yet after cooling at 1025 °C/min it still crystallized on reheating at 1900 °C/min, with an exotherm from a labelled 84.20 °C and melting near 131 °C baird2013 R10, C25. In the 2010 screen, 9 of 11 class II glasses recrystallized on reheating at 20 °C/min, and class I-B glasses crystallized rapidly when reheated above Tg baird2010 R8, R15, C24. Among the 31 drugs, the 19 class III compounds had MCVs from 216 µm/s (nimesulide) to 0.02 µm/s (clotrimazole): 7 were fast growers, 4 intermediate and all 8 slow growers belonged to class III trasi2014a R1, C14. The 12 class I and II compounds had MCVs of 1.8–104 µm/s, none of them slow growers trasi2014a R1, C14. Nilutamide formed a glass on cooling at 15 °C/min that did not crystallize on reheating, although its seeded stable-form crystals grew at about 1.5 µm/s at 70 °C and about 38–40 µm/s at 100 °C trasi2014b R2, R3, R8. Acetaminophen, cited from a 2011 thesis, readily forms a glass even at 1 °C/min cooling yet always recrystallizes on heating trasi2012a R1. Bicalutamide is cited as crystallizing rapidly above Tg despite high glass-forming ability, and celecoxib samples were "quite hard to nucleate" yet easily made amorphous moseson2022b R26; mosqueragiraldo2014 R15.

papersystemmethod and conditionsresultids
baird20134 class I-B, 10 class IIrapid heat–cool DSCRcrit,cool 25 °C/min (I-B) or 5 °C/min and below (II), against Rcrit,heat above 1500–2000 °C/min (I-B) and 12.5 to above 1910 °C/min (II)R9, C22, C24
baird2013flufenamic acidcooled at 25 or 1025 °C/minglass transition on reheating; crystallization on reheating even at 1900 °C/minR10, C25
baird201011 class II; 8 class I-BDSC reheating; Raman9 of 11 class II glasses recrystallized at 20 °C/min; class I-B glasses crystallized rapidly above TgR8, R15, C24, C47
trasi2014a19 class III and 12 class I/II drugsseeded melt growthclass III MCV 216 to 0.02 µm/s; class I/II 1.8–104 µm/sR1, C14, C15
trasi2014bnilutamideDSC; seeded growthglass on cooling without recrystallization on reheating; growth ≈38–40 µm/s at 100 °CR2, R8, C9
trasi2012aacetaminophencited DSC dataforms a glass even at slow cooling, always recrystallizes on heatingR1
moseson2022bbicalutamidecitedcrystallizes rapidly above Tg despite high glass-forming abilityR26
mosqueragiraldo2014celecoxibhot-stage microscopysamples hard to nucleate; easily rendered amorphous from the meltR15, C7

Figure

Fig. 2 of baird2013: flufenamic acid forms a glass at 25 °C/min yet crystallizes on reheating at 1900 °C/min
Fig. 2 · baird2013flufenamic acid forms a glass at 25 °C/min yet crystallizes on reheating at 1900 °C/min

Panel (a) has no exotherm on cooling and a glass transition on reheating; panel (b), after cooling at 1025 °C/min, still shows an exotherm and a melting endotherm on reheating at 1900 °C/min baird2013 R10.

Fig. 4 of trasi2014a: extended classification by nucleation and growth
Fig. 4 · trasi2014aextended classification by nucleation and growth

Growth runs upward and nucleation to the left; the box for rapid growth with low nucleation (class II, for example nimesulide, nilutamide, celecoxib) holds good glass formers that are at risk once nuclei exist; it is a schematic trasi2014a C17.

Why

In the authors' reading, a melt that forms a glass easily has nucleation and growth windows that overlap little: on cooling, the melt passes the growth maximum while few nuclei exist and forms nuclei only where growth is slow baird2013 C35, C36. On reheating, those nuclei, together with any formed on the way up, are carried into the fast-growth range, where a rate that sufficed on cooling is too slow; "assuming steady-state nucleation" as many nuclei form on reheating as on cooling, and the asymmetry arises because the reheating nuclei meet higher growth rates baird2013 C36. Nuclei formed during cooling are a further contribution (see F16) baird2013 C30, C40. For class III fast growers, Trasi and co-workers argue that ready glass formation can come from low nucleation, low growth, both, or poor overlap, and that the fast-growing class III compounds "clearly" form glasses because they nucleate poorly under the test conditions; nucleation rates were not measured trasi2014a C13, C15. For nilutamide, the authors attribute the glass-forming ability to a low nucleation tendency, since its crystals grow fast ("it appears") trasi2014b C9. Celecoxib's ease of amorphization is likewise attributed "most likely" to a low nucleation rate mosqueragiraldo2014 C7. Trasi and co-workers proposed an extended classification that separates these cases: IA (overlapping fast nucleation and growth, for example carbamazepine), IB (fast nucleation and growth with little overlap, acetaminophen), II (fast growth, low nucleation: nimesulide, nilutamide, celecoxib), III (ready nucleation, slow growth: cinnarizine, droperidol) and IV (slow nucleation and growth: warfarin, miconazole) trasi2014a C17. (synthesis: the 2010 classes rank the cooling step only, so two drugs in the same class III can differ by four orders of magnitude in how fast a nucleus, once formed, will grow; the growth rate is the missing measurement trasi2014a R1, C2.)

Conditions and limits

Neat drugs; DSC scans lasting minutes and growth measured above Tg baird2013 X15; trasi2014a P1. The low nucleation of fast-growing class III drugs is inferred, not measured trasi2014a C15. The 2014 MCVs carry no replicates and several do not match the plotted rates trasi2014a X17, X3. The class labels in the 2014 paper mix two schemes trasi2014a X10. Reheating rates of 1900–2000 °C/min exceed the calibrated range of the rapid calorimeter baird2013 X10.

Counterexamples and conflicts

In a small storage test the class III glasses were the most stable and the class II glasses less so, which fits the reheating results (see F17) baird2010 R24, R25, C47. Under 1–4 h isothermal holds of the undercooled melt, indomethacin (class III) did not nucleate while posaconazole and bicalutamide (also class III) crystallized within 1 h, so class III glasses differ widely in stability above Tg moseson2020 R7, C17, C18.

Transfer check

Always reheat the glass and measure a seeded growth rate; a clean cooling trace is not evidence of stability baird2010 C47; trasi2014a C16. A drug that forms a glass at slow cooling but crystallizes on reheating at 10–20 °C/min (class II), or that grows faster than about 10 µm/s at its maximum, should be treated as at risk once nuclei are present, for example after milling or incomplete melting (synthesis: from the class II and extended class II definitions) baird2010 R1; trasi2014a C16, C17. Where a fast-scanning calorimeter is available, compare Rcrit,cool with Rcrit,heat baird2013 C24.

History

Baird in 2010 separated glass-forming ability from glass stability and found class II much less stable than class III baird2010 C47. Acetaminophen illustrated the point in 2012 trasi2012a R1. Baird in 2013 quantified the cooling–heating asymmetry, reported as a first for organic systems baird2013 C26. Trasi in 2014 showed fast growers among good glass formers, proposed the extended classification, and found nilutamide a nucleation-limited glass former trasi2014a C14, C17; trasi2014b C9. Bicalutamide was cited as a further case in 2022 moseson2022b R26.

F16

Topic 3Distinction between forming a glass and keeping it2 of 3

Cooling a melt faster makes it crystallize less on reheating, which the authors attribute to fewer nuclei trapped in the glass.

Cooling at the maximum rate (above 500 °C/min) instead of 20 °C/min lowered the reheating rate needed to avoid crystallization in all ten comparable cases, slightly for tolazamide and more than three-fold for dibucaine baird2013 R9, C28. The authors read this as nuclei trapped in the glass during slower cooling; the nuclei themselves were never observed baird2013 C30, X7.

weakWeak to moderate: the cooling-rate effect is shown in thermograms, but the nuclei are inferred from rate effects and were not seen or counted baird2013 K5, K6, X7; baird2010 K8.

glass made by cooling at 20 °C/minglass made by cooling at the maximum ratedibucaine300above 1000nifedipine250above 500clofoctol200400cinnarizine135300droperidol125175tolazamide30351001000slowest reheating that avoids crystallization, °C/min (log scale)

Six of the ten compounds; the faster the glass was cooled, the lower the reheating rate that avoided crystallization. baird2013 R9, C28

For your drug

  1. In melt processes such as extrusion or melt quenching, record and control the cooling path, and test reheating and storage of material cooled as in the process, because the cooling rate changed the reheating behaviour of the same compound.
  2. Do not take a clean DSC cooling trace as evidence of a nucleus-free glass; run a storage study at the product's temperature before relying on the effect.

baird2013 R9, C28, X15, C41; baird2010 C54

How it works and key numbers
  1. The 2010 puzzle: class II melts form a glass yet crystallize on reheating. The explanation: nuclei form during cooling, cannot grow before the glass forms, and grow on reheating baird2010 C51, C55.
  2. On cooling at a moderate rate the melt first crosses the growth maximum while few nuclei exist, then forms nuclei near Tg where growth is slow and viscosity is high, so the nuclei stay in the glass baird2013 C36.
  3. On reheating, these nuclei enter the fast-growth range, where a rate that sufficed on cooling is too slow to stop growth baird2013 C36.
  4. Rapid cooling shortens the time in the undercooled range. If it gives a nucleus-free glass, crystallization on reheating needs both nucleation and growth, and the critical heating rate falls baird2013 C38, C28.
  5. A test with clofoctol: the cooling was identical (20 °C/min), only the reheating rate changed, yet the melting endotherm shrank from clear at 200 to absent at 500 °C/min, which the authors take to "confirm" that crystallization occurred on reheating baird2013 R12, C29.
  6. Not measured: the nuclei were not seen or counted, and nucleation and growth rates were not measured against temperature baird2013 X7, C35.
Analogy. Slow cooling lets seeds settle into the glass; reheating is the spring in which they sprout.
above 500 vs 20 °C/minmaximum against standard cooling rate; fast cooling lowered the critical heating rate in all ten comparable casesbaird2013 R9, C28
above 1000 → 300 °C/mindibucaine: critical heating rate after 20 °C/min against after fast coolingbaird2013 R9, X8
35 → 30 °C/mintolazamide, a small drop; droperidol is also smallbaird2013 R9, X8

Connections to other chapters

Evidence: data, conditions and papers

Finding F16. Faster cooling of the melt lowers later crystallization on reheating, which the authors attribute to quenched-in nuclei

In short. Cooling the melt at the maximum rate (above 500 °C/min) instead of 20 °C/min lowered the heating rate needed to avoid crystallization on reheating in all ten comparable class I-B and II cases, slightly for tolazamide (35 to 30 °C/min) and more than three-fold for dibucaine (above 1000 to 300 °C/min) baird2013 R9, C28. A clofoctol glass made at a constant 20 °C/min lost its melting endotherm when reheated at 500 instead of 200 °C/min, so its crystallization happened on reheating baird2013 R12, C29. The authors read this as nuclei formed during slower cooling and trapped in the glass ("strongly suggests"), as proposed in 2010 for class II; the nuclei themselves were never observed baird2013 C30, X7; baird2010 C55, C58.

Type. observation in N systems (10 compounds) with an untested interpretation Strength. to moderate — the cooling-rate effect is shown in thermograms (baird2013 K5: moderate), but the nuclei are inferred from DSC rate effects (baird2013 K6: weak to moderate; baird2010 K8: weak).

How it was shown

Class II melts form a glass on cooling yet crystallize on reheating, which Baird and co-workers explained in 2010 by nuclei that form during cooling and grow on reheating baird2010 C51, C55. In 2013 they made glasses of four class I-B and ten class II molecules by cooling at 20 °C/min or at the maximum rate, and reheated each at a series of rates to record Rcrit,heat baird2013 P2, P4, X5. Clofoctol was also cooled at a constant 20 °C/min and reheated at 200–500 °C/min baird2013 P4, R12.

What the data show

Rcrit,heat after 20 °C/min against after maximum cooling was 300 against 135 °C/min for cinnarizine, 400 against 200 for clofoctol and above 1000 against 300 for dibucaine baird2013 R9. The clofoctol endotherm was clear at 200 and absent at 500 °C/min reheating baird2013 R12.

papersystem and conditionsresultids
baird201310 class I-B and II compounds; Rcrit,heat after 20 °C/min against maximum coolinge.g. cinnarizine 300 to 135; clofoctol 400 to 200; dibucaine above 1000 to 300; nifedipine above 500 to 250; tolazamide 35 to 30 °C/minbaird2013 R9, C28
baird2013flufenamic acid, tolbutamide, acetaminophenno change readable; reheating rates exceed the instrument rangebaird2013 R9
baird2013dibucaine; reheating at 300 °C/min after 20 or 800 °C/min coolingendotherm after 20 °C/min cooling, none after 800 °C/minbaird2013 R11
baird2013clofoctol; constant 20 °C/min cooling, reheating at 200–500 °C/minendotherm clear at 200, absent at 500 °C/minbaird2013 R12, C29
baird2010class II and quench-glassy class Inuclei formed on cooling are the "most likely" cause of recrystallization on reheatingbaird2010 C55, C57, C58
Fig. 4 of baird2013: clofoctol reheating after 20 or 860 °C/min cooling, and after constant 20 °C/min cooling at four reheating rates
Fig. 4 · baird2013clofoctol reheating after 20 or 860 °C/min cooling, and after constant 20 °C/min cooling at four reheating rates

Panel (b) is the key test: the cooling was identical, and only the reheating rate changed, yet the endotherm shrinks from 200 to 500 °C/min baird2013 R12.

Why

On cooling at a moderate rate the melt crosses the growth maximum while few nuclei exist, then forms nuclei near Tg where growth is slow, so the nuclei stay in the glass and grow on reheating ("can account for") baird2013 C36. Reheating through Tg "should in principle" erase the cooling history, so a cooling-rate effect points to nuclei baird2013 C37. Nuclei were not seen or counted baird2013 X7.

Limits and transfer

The scans lasted minutes on neat drugs, and the highest reheating rates exceed the calibrated range of 10–1700 °C/min baird2013 A1, X15, X10. For droperidol and tolazamide the drop is small baird2013 X8, C4. Record and control the cooling path in melt processes, and do not take a clean DSC cooling trace as evidence of a nucleus-free glass baird2013 R9, C28; baird2010 C54.

How it was shown

The 2010 classification left a puzzle: class II melts form a glass on cooling yet crystallize on reheating, which Baird and co-workers explained by nuclei that form during cooling, cannot grow before the glass forms, and then grow on reheating baird2010 C51, C55. In 2013 the same group tested the idea with a prototype rapid heat/cool calorimeter on 30–50 µg crystals baird2013 P2. Glasses of four class I-B and ten class II molecules were made by cooling the melt either at 20 °C/min or at the maximum obtainable rate, stated as above 500 °C/min in all cases and printed as 1025 (flufenamic acid), 800 (dibucaine) and 860 °C/min (clofoctol) baird2013 P4, X5. Each glass was reheated at a series of rates, and the slowest reheating that avoided crystallization (a crystallization exotherm or a melting endotherm at Tm) was recorded as the critical heating rate Rcrit,heat baird2013 P4. To tell crystallization on reheating from crystallization on cooling, clofoctol was cooled at a constant 20 °C/min and reheated at 200, 300, 400 and 500 °C/min baird2013 P4, R12. Trasi and Taylor (2012) had approached the same question for acetaminophen by cooling melts to successively lower temperatures and reheating them in DSC and on a hot stage trasi2012a P6, P7, C40, X9.

What the data show

Rcrit,heat after 20 °C/min cooling against after maximum cooling was: antipyrin above 2000 against 2000, chlorpropamide above 2000 against 1000, cinnarizine 300 against 135, clofoctol 400 against 200, dibucaine above 1000 against 300, droperidol 175 against 125, nifedipine above 500 against 250, tolazamide 35 against 30, bifonazole 12.5 against below 10 and celecoxib 15 against below 10 °C/min baird2013 R9. No change could be read for flufenamic acid (above 1900 both), tolbutamide (above 1500 both) and acetaminophen (above 1910 both), and none was given for D(−) salicin baird2013 R9. The size of the drop therefore ranges from small (droperidol, tolazamide) to at least 3.3-fold (dibucaine) baird2013 X8. For dibucaine reheated at 300 °C/min, the glass made at 20 °C/min showed a melting endotherm near 66 °C and the glass made at 800 °C/min none; after 20 °C/min cooling the endotherm persisted, though shallower, at 300, 500 and 1000 °C/min (about 6, 3 and 1 W/g deep) baird2013 R11. For clofoctol reheated at 200 °C/min, a small endotherm near 89 °C appeared after 20 °C/min cooling and not after 860 °C/min cooling baird2013 R12. After a constant 20 °C/min cooling, the clofoctol endotherm was clear at 200, small at 300, at most barely visible at 400 and absent at 500 °C/min reheating baird2013 R12. Assuming that the same number of nuclei formed during the identical cooling, the authors take this to "confirm" that crystallization occurred during reheating baird2013 C29.

papersystemmethod and conditionsresultids
baird201310 class I-B and II compoundsRcrit,heat after 20 °C/min against maximum coolinge.g. cinnarizine 300 to 135; clofoctol 400 to 200; dibucaine above 1000 to 300; nifedipine above 500 to 250; tolazamide 35 to 30 °C/minR9, C28
baird2013flufenamic acid, tolbutamide, acetaminophenas aboveno change readable; reheating rates exceed the instrument rangeR9
baird2013dibucainereheating at 300 °C/min after 20 or 800 °C/min coolingendotherm after 20 °C/min cooling, none after 800 °C/minR11
baird2013clofoctolconstant 20 °C/min cooling, reheating at 200–500 °C/minendotherm clear at 200, absent at 500 °C/minR12, C29
baird2010class II and quench-glassy class Iinterpretation of the DSC classesnuclei formed on cooling are the "most likely" cause of recrystallization on reheating and are "believed" to lower stability below TgC55, C57, C58
trasi2012aacetaminopheninterpretation of cooling–reheating DSCquenched-in nuclei suggested; nuclei from cooling not separated from those formed on reheatingC40, X9

Figure

Fig. 3 of baird2013: dibucaine reheating thermograms after cooling at 20 or 800 °C/min
Fig. 3 · baird2013dibucaine reheating thermograms after cooling at 20 or 800 °C/min

In panel (a), at the same 300 °C/min reheating, the melting endotherm near 66 °C appears on the trace of the glass made at 20 °C/min (black) but not on the one made at 800 °C/min (red); panel (b) shows that after slow cooling even 1000 °C/min reheating leaves an endotherm baird2013 R11.

Fig. 4 of baird2013: clofoctol reheating after 20 or 860 °C/min cooling, and after constant 20 °C/min cooling at four reheating rates
Fig. 4 · baird2013clofoctol reheating after 20 or 860 °C/min cooling, and after constant 20 °C/min cooling at four reheating rates

Panel (b) is the key test: the cooling was identical, and only the reheating rate changed, yet the endotherm shrinks from 200 to 500 °C/min baird2013 R12.

Why

The authors' explanation starts from two rate curves with maxima at different temperatures, nucleation near Tg and growth closer to Tm baird2013 C35, C36. On cooling at a moderate rate the melt first crosses the growth maximum while few nuclei exist, then forms nuclei near Tg where growth is slow and viscosity high, so the nuclei stay in the glass baird2013 C36. On reheating these nuclei, plus any formed on reheating, enter the fast-growth range, where a rate that sufficed on cooling is too slow to stop growth ("can account for") baird2013 C36. Reheating through Tg "should in principle" erase the cooling history of the glass itself, so a cooling-rate effect on reheating points to something carried over, which they identify as nuclei baird2013 C37. Rapid cooling shortens the time in the undercooled range and so, "assuming" it gives a nucleus-free glass, crystallization on reheating then needs both nucleation and growth, and Rcrit,heat falls baird2013 C38, C28. From the smaller asymmetry after quenching they conclude that this "strongly suggests" nuclei generated during slower cooling, and in the conclusions "there is evidence" for quenched-in nuclei that "likely contribute" to crystallization on reheating baird2013 C30, C40. The 2010 paper had proposed the same nuclei, adding that below Tg, as the glass relaxes, molecules diffuse to them so that they seed crystal growth over long times ("It is believed") baird2010 C57, C58. A 2012 library paper restated that quenched-in nuclei may lead to observable crystallization over longer times trasi2012a C41. What was not measured: nuclei were not seen or counted, nucleation and growth rates were not measured as functions of temperature, and the clofoctol test rests on the stated assumption of equal nucleation during constant cooling baird2013 X7, C35. (synthesis: the glass made by fast cooling also differs in its structural state, which the authors set aside by assuming that reheating through Tg erases it baird2013 C37; a direct count of nuclei, for example by reheating into the growth range and counting crystals, would test the explanation.) The formula behind the argument is on the critical cooling rate to avoid nucleation page.

Conditions and limits

DSC scans lasting minutes on neat drugs, no polymer, no storage baird2013 A1, X15. The quench rate before reheating is printed for three compounds only baird2013 X5. The highest reheating rates (1900–2000 °C/min) exceed the calibrated range of 10–1700 °C/min, and replicate numbers are not stated baird2013 X10, X9. The clofoctol value in Table 4 (400 °C/min) conflicts with the text and figure, which place the threshold near 500 °C/min baird2013 X14.

Counterexamples and conflicts

No change could be read for flufenamic acid, tolbutamide and acetaminophen, whose reheating rates exceeded the instrument range baird2013 R9. The abstract's "greatly reduced" asymmetry holds for some compounds only; for droperidol and tolazamide the drop is small baird2013 X8, C4. In the acetaminophen protocol of 2012 the nucleation zone is passed both on cooling and on reheating, so nuclei formed on cooling were not separated from those formed on reheating trasi2012a X9.

Transfer check

In melt processes such as melt extrusion or melt quenching, record and control the cooling path, and test reheating and storage of material cooled as in the process, because the cooling rate changed the reheating behaviour of the same compound baird2013 R9, C28. Do not take a clean DSC cooling trace as evidence of a nucleus-free glass baird2010 C54. Storage effects of cooling history were not tested in these papers, so a storage study at the product's temperature is needed before relying on the effect baird2013 X15, C41.

History

Quenched-in nuclei were proposed in 2010 to explain the class II behaviour, with a TTT scheme adapted from Angell baird2010 C51, C58. Trasi and Taylor restated the idea for acetaminophen in 2012 trasi2012a C40. Baird in 2013 supported it indirectly with the cooling-rate effect on reheating and the clofoctol constant-cooling test baird2013 C30, C29.

F17

Topic 3Distinction between forming a glass and keeping it3 of 3

A drug's neat class is a first warning of how hard it will be to keep amorphous, not a forecast.

In a storage test of quenched glasses, indoprofen (class I) crystallized fully within a day, droperidol and nifedipine (class II) were fully crystalline after 3 months, and clotrimazole and felodipine (class III) crystallized only at the edges baird2010 A6, R23, R24, R25. With polymers, class I drugs needed more polymer than class II drugs vaneerdenbrugh2010b R4, C1, C15. But the link broke for some drugs, for example where crystalline PEG nucleated slowly crystallizing drugs zhu2012 C41; nguyen2023a C62.

weakWeak to moderate: the storage evidence is one or two compounds per class judged by eye, and the polymer screen is consistent but evaporation-based baird2010 K7, X16; vaneerdenbrugh2010b C1.

Fully crystalline within 3 months
indoprofendroperidolnifedipine
Only the edges crystallized at 3 months
clotrimazolefelodipine

Quenched glasses stored 14–20 °C below Tg at 0% RH; indoprofen (class I) was fully crystalline within a day, class II after 3 months, class III only at the edges. baird2010 R23, R24, R25

Class I drugsfive drugs47–59Class I/II drug68Class II drugstwo drugs73–97405060708090100average amorphous index of films with polymers, % (higher = easier to keep amorphous)

Spin-coated films of eight drugs with seven polymers, scored over 7 days. vaneerdenbrugh2010b R4, C15

For your drug

  1. Treat class I as a warning that strong crystallization inhibition will be needed, and class II as a warning that the glass may carry nuclei.
  2. Then test the actual dispersion at its drug loading, polymer, humidity, temperature and particle surface, because the class alone did not predict pretomanid, delamanid, PEG or tacrolimus outcomes. With a crystalline carrier such as PEG, test whether the carrier nucleates the drug.

vaneerdenbrugh2010a C48; baird2010 C58; nguyen2023a C62; duong2022a C44; zhu2012 C41, C4; trasi2017 C28

How it works and key numbers
  1. The authors attribute the faster crystallization of class II and quench-glassy class I glasses below Tg to nuclei formed during cooling, to which molecules diffuse as the glass relaxes ("It is believed"). The nuclei were not detected baird2010 C57, C58.
  2. For dispersions, the polymer screen reads the class as how much inhibition a drug needs; class I drugs "may not be appropriate candidates" for solid dispersions vaneerdenbrugh2010b C1; vaneerdenbrugh2010a C48.
  3. Where the link breaks: both the drug's own tendency and its ability to interact with the carrier matter, and a crystalline carrier can add heterogeneous nucleation zhu2012 C38, C39, C4.
  4. Pretomanid (class I) glass crystallized within minutes, yet its HPMCAS dispersions were stable for several months; a commercial tacrolimus product crystallized faster than the pure drug nguyen2023a R4, R27, C62; trasi2017 R16, C28.
  5. The extractor's reading: the class is measured on the neat drug over short times, while stability in a product also depends on polymer, water, surfaces and process history zhu2012 C41; nguyen2023a C62; trasi2017 C28.
47–59% vs 73–97%average amorphous index with polymers, class I drugs against class II drugsvaneerdenbrugh2010b R4, C15
1 day vs 3 monthsindoprofen (class I) fully crystalline after a day; class II drugs only after 3 monthsbaird2010 R23, R24
minutes vs monthspretomanid (class I): neat glass crystallized in minutes, HPMCAS dispersion stayed stable for monthsnguyen2023a R4, R27, C62

Connections to other chapters

10 more connections
Evidence: data, conditions and papers

Finding F17. The neat-drug class is a first risk flag for keeping a drug amorphous, in storage and in polymer dispersions, not a prediction

In short. Neat-drug crystallization tendency ranked how hard drugs were to keep amorphous, but only qualitatively baird2010 C46; vaneerdenbrugh2010b C1. In a storage test of one or two quenched glasses per class (0% RH, 14–20 °C below Tg), indoprofen (class I) crystallized fully within a day, droperidol and nifedipine (class II) appeared fully crystalline after 3 months, and clotrimazole and felodipine (class III) crystallized only at the edges baird2010 A6, R23, R24, R25, C46. With polymers, the class I drugs of a spin-coat screen still had lower amorphous indices than the class II drugs and needed more polymer vaneerdenbrugh2010b R4, C1, C15. The link broke where crystalline PEG nucleated slowly crystallizing drugs, and an HPMCAS dispersion of class I pretomanid stayed amorphous for months zhu2012 C41; nguyen2023a C62.

Type. observation in N systems (5 stored glasses; 8 drugs with polymers; several later drugs) Strength. to moderate — the storage evidence is one or two compounds per class judged by eye (baird2010 K7: weak to moderate; X16), and the polymer screen is consistent but evaporation-based (vaneerdenbrugh2010b C1).

How it was shown

Baird and co-workers (2010) quenched melts of five molecules in liquid N2 between two cover glasses and stored them over P2O5 (0% RH), 14–20 °C below Tg, following crystallization by polarized light microscopy baird2010 P7, A6, C45, X16. Van Eerdenbrugh and Taylor (2010) spin coated eight drugs of known class with seven polymers and averaged an amorphous index per drug over 7 days of dry storage vaneerdenbrugh2010b P1, P2, R2, E4.

What the data show

Indoprofen showed one spherulite after 5 min and a fully birefringent field after 1 day baird2010 R23. Clotrimazole and felodipine showed only edge crystallization after 3 months baird2010 R25. Class I drugs averaged 47–59% amorphous index and class II drugs 73–97% vaneerdenbrugh2010b R4, C15.

papersystemmethod and conditionsresultids
baird2010indoprofen; droperidol, nifedipine; clotrimazole, felodipinequenched glasses between cover glasses, 0% RH, 15 or 30 °Ccomplete crystallization within 1 day; complete by 3 months; edges only at 3 monthsbaird2010 R23, R24, R25, X16
vaneerdenbrugh2010b8 drugs with 7 polymersspin-coated films, 7 days dryaverage amorphous index 47–59% for class I drugs, 68% for the class I/II drug, 73–97% for class II drugsvaneerdenbrugh2010b R4, C1, C15
zhu2012ibuprofen, fenofibrate in PEG25 °Clink between pure-drug and dispersion crystallization breaks down because crystalline PEG promotes heterogeneous nucleationzhu2012 C4, C41
nguyen2023apretomanid (class I)HPMCAS dispersions at ambient conditionsstable for several months; neat glass crystallized within minutesnguyen2023a R4, R27, C62
trasi2017tacrolimus40 °C/75% RH, open dishpure amorphous drug starts to crystallize after 2 weeks; a commercial product faster than the pure drugtrasi2017 R16, C28
Fig. 4 of vaneerdenbrugh2010b: average amorphous index per drug across seven polymers
Fig. 4 · vaneerdenbrugh2010baverage amorphous index per drug across seven polymers

The class I drugs sit lowest and the class II drugs highest, the same order as their neat-drug classes vaneerdenbrugh2010b R4.

Why

Baird and co-workers attribute the faster crystallization of class II and quench-glassy class I glasses below Tg to nuclei formed during cooling ("It is believed"); the nuclei were not detected baird2010 C57, C58. Zhu and co-workers explain where the link breaks: both the drug's own tendency and its ability to interact with the carrier matter, and a crystalline carrier can add heterogeneous nucleation zhu2012 C38, C39, C4.

Limits and transfer

The storage ranking rests on one or two compounds per class, one temperature each, judged by eye, at 0% RH only baird2010 X16, P7, A6. Treat class I as a warning that strong inhibition will be needed, then test the actual dispersion at its drug loading, polymer, humidity and temperature, because the class alone did not predict pretomanid, delamanid, PEG or tacrolimus outcomes vaneerdenbrugh2010a C48; nguyen2023a C62; duong2022a C44; zhu2012 C41; trasi2017 C28.

How it was shown

Baird and co-workers (2010) quenched melts of five molecules in liquid N2 between two cover glasses, which they assumed minimizes surface crystallization, and stored them over P2O5 (0% RH): indoprofen at 30 °C, droperidol and clotrimazole at 15 °C, nifedipine and felodipine at 30 °C, that is 14–20 °C below Tg baird2010 P7, A6, C45. Crystallization was followed by polarized light microscopy at 5 min, 1 day and 3 months, without a crystallinity measurement baird2010 P7, X16. Van Eerdenbrugh and Taylor (2010) spin coated eight drugs of known class with seven polymers at seven drug/polymer ratios from ethanol, scored the films by polarized light at days 0, 1, 3 and 7 of dry storage, and averaged an amorphous index per drug vaneerdenbrugh2010b P1, P2, R2, E4. Marsac and co-workers (2006) had compared nucleation in pure films of nifedipine and felodipine and in their PVP dispersions marsac2006a R1, C52. Zhu and co-workers (2012) measured crystallization of six drugs in 20/80 PEG dispersions against the pure melts at 25 °C, using 50 samples per pure drug for induction-time statistics zhu2012 R1, R2, R3, R23. Later papers used the neat-drug class to choose model drugs or to explain dispersion results for delamanid, pretomanid, lumefantrine, tacrolimus and three melt-extrusion drugs duong2022a C44; nguyen2023a C59; trasi2020 C7; trasi2017 C6; moseson2020 C5.

What the data show

Quenched indoprofen (Tg 50 °C) stored at 30 °C showed one spherulite after 5 min and a fully birefringent field after 1 day baird2010 R23. Droperidol at 15 °C (Tg 29 °C) and nifedipine at 30 °C (Tg 45 °C) crystallized at the edges within 1 day and appeared completely crystalline after 3 months baird2010 R24. Clotrimazole at 15 °C (Tg 30 °C) and felodipine at 30 °C (Tg 45 °C) showed only edge crystallization after 3 months baird2010 R25. The authors conclude "there may be a link" between glass-forming ability and glass stability baird2010 C46, C47. In the polymer screen, average amorphous indices were 47–59% for the five class I drugs, 68% for the class I/II drug and 73–97% for the two class II drugs vaneerdenbrugh2010b R4, C15. The authors conclude that drugs whose pure form crystallizes more readily were harder to stabilize and needed more polymer vaneerdenbrugh2010b C1. Pure nifedipine nucleated faster than felodipine (0.39 against 0.17 × 10¹⁰ m⁻³ day⁻¹ at 22 °C), and the authors suggest that the relative tendency in dispersions "may be dependent" on that of the pure drugs marsac2006a R4.a, C52. In PEG, haloperidol, which crystallizes within seconds as a pure melt, was hardly affected, and benzocaine was slowed by its favourable interaction with PEG zhu2012 R1, C39. Ibuprofen and fenofibrate, which as pure liquids started to crystallize in only 2 and 4 of 50 samples within 1 h, nucleated more readily in the dispersion, with crystals forming at crystalline PEG surfaces zhu2012 R2, R3, C4, C34. The authors state that the link between pure-drug and dispersion crystallization "was found to break down" for these slowly crystallizing drugs zhu2012 C41. Pretomanid (class I) glass crystallized within minutes, yet its HPMCAS dispersions were physically stable under ambient conditions for several months nguyen2023a R4, R27, C62. Delamanid (class I) dispersions made by rotary evaporation held crystals above about 10–15% drug loading duong2022a R8, C44. Pure amorphous tacrolimus did not start to crystallize until after 2 weeks at 40 °C/75% RH, while a commercial formulation crystallized faster than the pure drug, pointing to a formulation or processing factor trasi2017 R16, C28. Under 1–4 h holds of the undercooled melt, posaconazole and bicalutamide (both class III) crystallized within 1 h while indomethacin did not nucleate in 4 h moseson2020 R7, C17, C18.

papersystemmethod and conditionsresultids
baird2010indoprofen; droperidol, nifedipine; clotrimazole, felodipinequenched glasses between cover glasses, 0% RH, 15 or 30 °Ccomplete crystallization within 1 day; complete by 3 months; edges only at 3 monthsR23, R24, R25, X16
vaneerdenbrugh2010b8 drugs with 7 polymersspin-coated films, 7 days dryaverage amorphous index 47–59% for class I drugs, 68% for the class I/II drug, 73–97% for class II drugsR4, C1, C15, C16
marsac2006anifedipine, felodipinepure films at 22 °C, 0% RH; dispersionsnifedipine nucleates faster; the relative tendency in dispersions "may" follow that of the pure drugsR1, R4, C52
zhu2012ibuprofen, fenofibrate in PEG25 °Clink between pure-drug and dispersion crystallization breaks down because crystalline PEG promotes heterogeneous nucleationC4, C41
duong2022adelamanid (class I)rotary-evaporated dispersionscrystal-free dispersions above about 10% drug loading difficult to prepareR8, C44
nguyen2023apretomanid (class I)HPMCAS dispersions at ambient conditionsstable for several months; neat glass crystallized within minutesR4, R27, C62
trasi2017tacrolimus40 °C/75% RH, open dishpure amorphous drug starts to crystallize after 2 weeks; a commercial product faster than the pure drugR16, C28
moseson2020posaconazole, bicalutamide, indomethacin (all class III)isothermal holds of the undercooled melt, 130–180 °Cposaconazole and bicalutamide crystallized within 1 h; indomethacin did not nucleate in 4 hR7, C17, C18
trasi2020lumefantrine (class 3)dispersion granulesgood glass-forming ability makes it a good dispersion candidate ("suggesting")C27

Figure

Fig. 5 of baird2010: quenched glasses stored below Tg, under crossed polarizers
Fig. 5 · baird2010quenched glasses stored below Tg, under crossed polarizers

Bright areas are crystals and a dark field is glass: indoprofen at 5 min and 1 day (a, b), droperidol and clotrimazole at 15 °C after 3 months (c, d), nifedipine and felodipine at 30 °C after 3 months (e, f); the extractor notes that panels (c) and (d) do not visibly match the text baird2010 R23, R24, R25, X16.

Fig. 4 of vaneerdenbrugh2010b: average amorphous index per drug across seven polymers
Fig. 4 · vaneerdenbrugh2010baverage amorphous index per drug across seven polymers

The class I drugs sit lowest and the class II drugs highest, the same order as their neat-drug classes vaneerdenbrugh2010b R4.

Why

Baird and co-workers attribute the faster crystallization of class II and quench-glassy class I glasses below Tg to nuclei formed during cooling, to which molecules diffuse as the glass relaxes ("It is believed"); the nuclei were not detected baird2010 C57, C58. Indoprofen's rapid crystallization below Tg is read as a high inherent tendency from the glass baird2010 C44. For dispersions, the 2010 polymer screen reads the class as how much inhibition a drug needs, and Van Eerdenbrugh and co-workers write that class I drugs "may not be appropriate candidates" for solid dispersions vaneerdenbrugh2010b C1; vaneerdenbrugh2010a C48. Zhu and co-workers explain where the link breaks: both the drug's own tendency and its ability to interact with the carrier matter, and a crystalline carrier can add heterogeneous nucleation zhu2012 C38, C39, C4. Trasi and co-workers grade the risk by the extended classes, class I highest, then class II, class III intermediate and class IV "likely" the most robust, assuming relative growth rates carry over to storage temperatures trasi2014a C18, C16. Tacrolimus authors name inherent crystallization tendency as the most important parameter of physical instability, set by conformational flexibility, configurational entropy, driving force, viscosity and hydrogen bonding, "not fully elucidated" trasi2017 C6. (synthesis: the class is measured on the neat drug at short time scales, whereas stability in a product depends also on the polymer, water, surfaces and process history; the cases above where the class failed involve exactly one of these factors: a nucleating carrier, a strong inhibitor, or a formulation step zhu2012 C41; nguyen2023a C62; trasi2017 C28.)

Conditions and limits

The storage ranking rests on one or two compounds per class, one temperature each, judged by eye, and indoprofen is really class I-B by its quench data baird2010 X16, X3. Only 0% RH and sandwiched samples were used, so humidity and free surfaces were not examined (see F21) baird2010 P7, A6, C45. The polymer screen used spin-coated films over 7 days vaneerdenbrugh2010b P1, P2. The later dispersion results come from single drugs and different preparation routes duong2022a C44; nguyen2023a R27.

Counterexamples and conflicts

PEG dispersions of slowly crystallizing drugs zhu2012 C41. Pretomanid dispersions, class I yet stable for months with HPMCAS nguyen2023a C62. Class III drugs differ widely under isothermal holds of the melt moseson2020 R7. A commercial tacrolimus product crystallized faster than the pure amorphous drug trasi2017 C28.

Transfer check

Treat class I as a warning that strong crystallization inhibition will be needed, and class II as a warning that the glass may carry nuclei vaneerdenbrugh2010a C48; baird2010 C58. Then test the actual dispersion at its drug loading, polymer, humidity, temperature and particle surface, because the class alone did not predict pretomanid, delamanid, PEG or tacrolimus outcomes nguyen2023a C62; duong2022a C44; zhu2012 C41; trasi2017 C28. For a crystalline carrier such as PEG, test whether the carrier nucleates the drug zhu2012 C4.

History

Marsac in 2006 suggested that pure-drug tendency carries into dispersions marsac2006a C52. Baird and Van Eerdenbrugh in 2010 proposed the link from the classes and the polymer screen baird2010 C46; vaneerdenbrugh2010b C1. Zhu in 2012 found where it breaks zhu2012 C41. Trasi in 2014 graded the risk with growth rates trasi2014a C18. Dispersion work from 2017 to 2023 used the class to choose drugs and strategies and found further exceptions trasi2017 C6; moseson2020 C5; duong2022a C44; nguyen2023a C62.

Topic 4 of 4 · 4 findings · F18–F21

Formulation consequences

F18

Topic 4Formulation consequences1 of 4

How a drug crystallizes from its melt only partly predicts how it crystallizes from a supersaturated water solution.

Some orderings carried over: felodipine, slow from the melt, underwent liquid–liquid phase separation (formation of a drug-rich liquid phase) in water, while faster-crystallizing nifedipine crystallized just below the coexistence curve raina2014a R20, C31. But several drugs classed as slow from the melt, such as efavirenz, papaverine, warfarin and glyburide, crystallized readily from supersaturated solution raina2015b C13, C28; ilevbare2013c R8, C24; hsieh2014 R18, C30.

moderateModerate: several independent comparisons point the same way, but each has few compounds and the solid-state classes are mostly cited from the 2010 papers raina2015b R1, P6; ilevbare2013c R8; hsieh2012 P7.

First time the precipitate was crystalline in water, hnifedipinemelt class II0.25nimodipinemelt class II3nitrendipinemelt class III3felodipinemelt class III6nisoldipinemelt class III6cilnidipinemelt class III12

Six dihydropyridines; nitrendipine, class III from the melt, crystallized from solution as early as class II nimodipine. raina2015b R1, R6

Induction time without polymer, min (log scale)celecoxibmelt class IIabout 2efavirenzmelt class III5ritonavirmelt class III120

In pH 6.8 buffer at 37 °C. Two class III drugs differ by more than an order of magnitude, and efavirenz is nearly as fast as celecoxib. ilevbare2013c R8, C24

For your drug

  1. Measure induction time and the form of the precipitate in the medium of interest, at the supersaturation the formulation will create, and with and without the intended polymer; do not infer solution behaviour from the DSC class.
  2. A class I drug is likely to crystallize fast from solution as well, but a class III drug may or may not.

ilevbare2013c R8; hsieh2014 C30, R18; raina2015b C28; nguyen2023a C59

How it works and key numbers
  1. Slow crystallization "may explain" why slow crystallizers undergo liquid–liquid phase separation first; if nucleation is fast relative to that separation, it is not seen ilevbare2013a C27; raina2014a C30, C31.
  2. Celecoxib and efavirenz nucleate much faster than ritonavir ("suggests"); ritonavir's slow nucleation is attributed "most likely" to its higher molecular weight and conformational flexibility ilevbare2013c C24, C25.
  3. Molecules with several low-energy forms, such as glyburide and warfarin with their tautomers, may crystallize more slowly because the effective supersaturation of the right form is reduced ("may explain") hsieh2014 C31.
  4. The extractor's reading: the melt classes compress a continuous property into three bins at fixed DSC rates (see F10), while water adds the medium, the supersaturation and the drug-rich phase, so a match at the extremes and a mismatch within class III are both to be expected raina2015b C28; hsieh2012 C42, X1.
0.25 h vs 12 hfirst sampling time with crystals in water, nifedipine against cilnidipineraina2015b R6
2, 5 and 120 mininduction times of celecoxib, efavirenz and ritonavir without polymerilevbare2013c R8
4 of 6dihydropyridines in one "slow" class from the melt, spread over four classes in solutionraina2015b C28

Connections to other chapters

Evidence: data, conditions and papers

Finding F18. Solid-state crystallization tendency predicts crystallization from supersaturated aqueous solution only partly

In short. Some orderings carried over: felodipine, slow from the melt, exceeded its amorphous solubility and underwent liquid–liquid phase separation in water, while faster-crystallizing nifedipine crystallized just below the coexistence curve raina2014a R20, C31; raina2014b C28. But aqueous rankings discriminated much more finely than the melt classes, and several compounds classed as slow crystallizers from the melt crystallized readily from supersaturated solution, such as efavirenz, papaverine, warfarin and glyburide raina2015b C13, C28; ilevbare2013c R8, C24; hsieh2014 R18, C30.

Type. observation in N systems (dihydropyridines, weak bases, ritonavir, efavirenz, celecoxib) Strength. several independent comparisons point the same way, each with few compounds, and the solid-state classes are mostly cited from the 2010 papers.

How it was shown

The classes come from the 2010 melt and spin-coat screens hsieh2012 P7; ilevbare2013c C24; raina2015b P6. Raina and co-workers (2015) precipitated six dihydropyridines by solvent shift into pH 6.8 buffer, sampled the precipitates by synchrotron WAXS at 0.25 to 24 h, and classed the same drugs by DSC raina2015b P1, P4, P6, R6. Ilevbare and co-workers (2013) measured induction times of ritonavir, efavirenz and celecoxib without polymer in pH 6.8 buffer at 37 °C ilevbare2013c R8.

What the data show

By DSC, felodipine, cilnidipine, nisoldipine and nitrendipine were class III and nifedipine and nimodipine class II raina2015b R1, C12. In water the first sampling time with crystallinity was 0.25 h for nifedipine and 12 h for cilnidipine, so solution discriminates "considerably" more than the melt raina2015b R6, C13, C28.

papersystemmethod and conditionsresultids
raina2015bsix dihydropyridinessolvent-shift precipitation, WAXS at 0.25–24 h, against DSC classsolution order cilnidipine < felodipine = nisoldipine < nitrendipine = nimodipine < nifedipine; only nimodipine and nifedipine recrystallize from meltsraina2015b R1, R6, C13, C28
ilevbare2013critonavir, efavirenz (class III), celecoxib (class II)induction time, pH 6.8 buffer, 37 °C, n = 3about 2 (celecoxib), 5 (efavirenz) and 120 min (ritonavir)ilevbare2013c R8, C24
hsieh2014papaverine, warfarin, glyburide (slow by melt quench)pH-induced precipitation, WAXSprecipitated crystalline, glyburide after about 15–30 min; the melt class "may not necessarily" predicthsieh2014 R18, C30
raina2014afelodipine, nifedipinesupersaturated solutionsfelodipine undergoes LLPS; nifedipine crystallizes below the coexistence curve without inhibitorraina2014a R20, C31
nguyen2023a; duong2022apretomanid, delamanid (class I)induction time near the amorphous solubilitycrystallize within minutes, matching the solid-state classnguyen2023a C59; duong2022a C10, R2
Fig. 10 of raina2015b: melt and solution classes of six dihydropyridines side by side
Fig. 10 · raina2015bmelt and solution classes of six dihydropyridines side by side

From the melt four of the six drugs fall in one "slow" class; in solution the same drugs spread over four classes raina2015b C28.

Why

Ilevbare and co-workers read the fast crystallization of celecoxib and efavirenz as much faster nucleation than ritonavir ("suggests"), and attribute ritonavir's slow nucleation "most likely" to its higher molecular weight and conformational flexibility ilevbare2013c C24, C25. Raina and co-workers relate the solution order to the competition between crystallization and LLPS raina2014a C30, C31.

Limits and transfer

Each paper used different compound sets, supersaturation methods and detection methods, with few compounds per comparison ilevbare2013c P9, R8; hsieh2014 R1, R4; raina2015b P1, P4, R1. Class I pretomanid and delamanid crystallize fast from solution too nguyen2023a C59; duong2022a C10. Measure induction time and precipitate form in the medium of interest and do not infer solution behaviour from the DSC class ilevbare2013c R8; hsieh2014 C30; raina2015b C28.

How it was shown

The classes in this finding come from the 2010 melt and spin-coat screens and were set against crystallization from water in later papers hsieh2012 P7; ilevbare2013c C24; raina2015b P6. Hsieh and co-workers (2012) generated supersaturation of ten weak bases by potentiometric pH titration (CheqSol) and sorted them into "type 1" (rapid precipitation) and "type 2" (long-lived supersaturation) compounds, then compared the types with each drug's class from the melt and from solvent evaporation hsieh2012 P7, M5, C42. Ilevbare and co-workers (2013) measured nucleation induction times of ritonavir, efavirenz and celecoxib without polymer in pH 6.8 phosphate buffer at 37 °C, in triplicate, and watched amorphous particles of each drug in buffer under polarized light ilevbare2013c R8, R4, R5. Hsieh and co-workers (2014) followed pH-induced precipitation of papaverine, dipyridamole, glyburide and warfarin by synchrotron WAXS hsieh2014 R4, R18. Raina and co-workers (2014) compared felodipine and nifedipine supersaturated solutions with and without polymer raina2014a R20, C31; raina2014b C27. Raina and co-workers (2015) precipitated six dihydropyridines by solvent shift (DMSO stock diluted 1:9 into pH 6.8 buffer, 10 mg/mL nominal) and sampled the precipitates by synchrotron WAXS at 0.25 to 24 h; they classed the same six drugs by DSC (cooling at 20 °C/min, reheating at 20 °C/min) raina2015b P1, P4, P6, R6.

What the data show

By DSC from the undercooled melt, felodipine, cilnidipine, nisoldipine and nitrendipine were class III and nifedipine and nimodipine class II raina2015b R1, C12. In water the same six spread out: the first sampling time with crystallinity was 0.25 h for nifedipine, 3 h for nimodipine and nitrendipine, 6 h for felodipine and nisoldipine, and 12 h for cilnidipine raina2015b R6. The authors give the order cilnidipine < felodipine = nisoldipine < nitrendipine = nimodipine < nifedipine and state that solution discriminates "considerably" more than the melt, where only nimodipine and nifedipine recrystallize raina2015b C13, C28. Nitrendipine, class III from the melt, thus crystallized from solution as early as class II nimodipine raina2015b R1, R6. Without polymer, induction times were about 2 min for celecoxib (class II), 5 min for efavirenz and 120 min for ritonavir (both class III) ilevbare2013c R8, C24. Melt-quenched celecoxib particles showed birefringent spots within about 1 min in buffer, while efavirenz particles shrank without birefringence over 3 h ilevbare2013c R4, R5. Papaverine, warfarin and glyburide, classed as slow crystallizers by melt quench, precipitated crystalline on rapid supersaturation, glyburide after about 15–30 min, so the melt class "may not necessarily" predict behaviour in dilute aqueous solution hsieh2014 R18, C30. In the 2012 titration study, all type 2 compounds were slow crystallizers, but class III also covered rapidly precipitating type 1 compounds, so the class did not separate the two types; dipyridamole is class I from the melt and class III on solvent evaporation hsieh2012 C42, X1, M5. Felodipine, a slow crystallizer, can exceed its amorphous solubility and undergo LLPS, whereas nifedipine, a faster crystallizer, crystallizes just below the coexistence curve without an inhibitor raina2014a R20, C31. Amorphous nifedipine and felodipine powders differed in crystallization in water only when polymers were present raina2014b C27. Class I pretomanid and delamanid both crystallized within minutes from solution near their amorphous solubility, in line with their solid-state class nguyen2023a C59; duong2022a C10, R2.

papersystemmethod and conditionsresultids
raina2015bsix dihydropyridinessolvent-shift precipitation, WAXS at 0.25–24 h, against DSC classsolution order cilnidipine < felodipine = nisoldipine < nitrendipine = nimodipine < nifedipine; only nimodipine and nifedipine recrystallize from meltsR1, R6, C13, C28
ilevbare2013critonavir, efavirenz (class III), celecoxib (class II)induction time, pH 6.8 buffer, 37 °C, n = 3about 2 (celecoxib), 5 (efavirenz) and 120 min (ritonavir)R8, C24
hsieh2014papaverine, warfarin, glyburide (slow by melt quench)pH-induced precipitation, WAXSprecipitated crystalline, glyburide after about 15–30 min; the melt class "may not necessarily" predictR18, C30
hsieh2012ten weakly basic compoundspH-induced precipitation (CheqSol)long-lived supersaturation only in slow crystallizers, correlation "to some extent"; class III also covers rapidly precipitating compoundsC42, X1, M5
raina2014afelodipine, nifedipinesupersaturated solutionsfelodipine undergoes LLPS; nifedipine crystallizes below the coexistence curve without inhibitorR20, C31
raina2014bnifedipine, felodipine powdersdissolution in aqueous mediadifferent crystallization tendencies show only with polymers; nifedipine's faster crystallization "seems" to carry overC27, C28
ilevbare2013aslow crystallizerssupersaturated solutionsslow crystallization "may explain" LLPS before crystallizationC27
nguyen2023a; duong2022apretomanid, delamanid (class I)induction time near the amorphous solubilitycrystallize within minutes, matching the solid-state classC59; C10, R2

Figure

Fig. 10 of raina2015b: melt and solution classes of six dihydropyridines side by side
Fig. 10 · raina2015bmelt and solution classes of six dihydropyridines side by side

From the melt four of the six drugs fall in one "slow" class; in solution the same drugs spread over four classes raina2015b C28.

Fig. 5 of raina2015b: when each precipitate turned crystalline, without polymer
Fig. 5 · raina2015bwhen each precipitate turned crystalline, without polymer

Grey cells are amorphous and black crystalline at 0.25–24 h; nifedipine is black from the first sample and cilnidipine only from 12 h raina2015b R6.

Why

Ilevbare and co-workers read the fast crystallization of celecoxib and efavirenz after amorphous dissolution as much faster nucleation than ritonavir ("suggests"), and attribute ritonavir's slow nucleation "most likely" to its higher molecular weight and conformational flexibility ilevbare2013c C24, C25. Hsieh and co-workers note that molecules with several low-energy forms, such as glyburide and warfarin with their tautomers, may crystallize more slowly because the effective supersaturation of the right form is reduced ("may explain") hsieh2014 C31. Raina and co-workers relate the solution order to the competition between crystallization and LLPS: if nucleation is fast relative to LLPS, phase separation is not seen raina2014a C30, C31. Slow crystallization "may explain" the tendency of slow crystallizers to undergo LLPS first ilevbare2013a C27. Nifedipine's faster inherent crystallization "seems" to carry over to water, complicated by a transient polymorph raina2014b C28. (synthesis: none of these papers measured the same drug's nucleation rate in the melt and in water, so the comparisons rest on classes and induction times.) (synthesis: the melt classes compress a continuous property into three bins at fixed DSC rates (see F10), while crystallization from water adds the medium, the supersaturation and the amorphous drug-rich phase, so a match at the extremes and a mismatch within class III are both to be expected.)

Conditions and limits

Aqueous media at 37 °C for the 2013 induction times and 25 °C for the 2014 pH-shift supersaturation, with no temperature stated for the 2015 solvent-shift study, and different compound sets, supersaturation methods and detection methods in each paper ilevbare2013c P9, R8; hsieh2014 R1, R4; raina2015b P1, P4. The solid-state classes are cited from the 2010 papers except for the 2015 DSC screen, which reheated at 20 instead of 10 °C/min raina2015b P6; hsieh2012 P7. Few compounds per comparison raina2015b R1; ilevbare2013c R8.

Counterexamples and conflicts

Felodipine and nifedipine keep the solid-state order in water raina2014a C31; raina2014b C28. Class I pretomanid and delamanid crystallize fast from solution as well nguyen2023a C59; duong2022a C10. Polymer rankings also change between solid state and solution: for acetaminophen, PAA was an excellent growth inhibitor in the supercooled liquid but largely ineffective in solution trasi2015 C26, C28.

Transfer check

Measure induction time and the form of the precipitate in the medium of interest, at the supersaturation the formulation will create, and with and without the intended polymer; do not infer solution behaviour from the DSC class ilevbare2013c R8; hsieh2014 C30; raina2015b C28. A class I drug is likely to crystallize fast from solution as well, but a class III drug may or may not nguyen2023a C59; hsieh2014 R18.

History

Hsieh in 2012 found a partial correlation between solution behaviour and solid-state class hsieh2012 C42. Ilevbare in 2013 and Hsieh in 2014 found melt slow crystallizers that crystallize fast in solution ilevbare2013c C24; hsieh2014 C30. Raina in 2014 linked the slow class with LLPS and in 2015 showed that solution discriminates more than the melt raina2014a C31; raina2015b C28.

F19

Topic 4Formulation consequences2 of 4

A polymer slows crystal growth by a similar factor in fast and slow growers, so fast growers need more polymer.

Adding 10 mol% PVP (a polymer) lowered the melt growth rates of five hydrogen-bond-donor drugs by roughly 3- to 14-fold at most temperatures, although their neat rates differed widely. The authors conclude that faster growers "will clearly require" more polymer for a given rate trasi2014a R6.m, R6.c, X6, C10, C11.

moderateModerate: measured at several temperatures per drug with consistent ratios for four of five drugs, but without replicates and on an undefined molar basis trasi2014a K5, X17, X7.

neat drugwith 10 mol% PVPflutamideat 70 °C5.678acetaminophenat 100 °C7.445indomethacinat 120 °C0.171.25aceclofenacat 50 °C0.00350.0280.0010.010.1110100melt growth rate, µm/s (log scale)

Read from the 2014 figure: the neat rates differ by orders of magnitude, but each rate falls by a similar factor with PVP. trasi2014a R6, X6

Nilutamide growth at 80 °C, µm/s: the polymer mattersno polymer≈510% molar PAA3.810% molar PVP1.1

PVP, the better hydrogen-bond acceptor, slowed growth far more than PAA. trasi2014b R3, C19

For your drug

  1. Measure the neat growth rate and the reduction factor of the candidate polymer at two temperatures, including one close to storage, and choose the loading for the rate needed.
  2. Check by FTIR on drug–polymer films that the drug has a hydrogen-bond donor and the polymer a good acceptor before expecting a PVP-like factor. Test growth and nucleation separately, because the best growth inhibitor can promote nucleation.

trasi2014a C11, C12; kestur2010b R13; trasi2014b C18; trasi2012a C46

How it works and key numbers
  1. The authors attribute growth inhibition mainly to specific drug–polymer interactions: for nilutamide the interaction strength "correlates well" with the inhibition, and PVP is a better hydrogen-bond acceptor than the drug itself trasi2014b C19, C18.
  2. For the five 2014 drugs, all good hydrogen-bond donors, they assume similar bonding with PVP ("would be expected"; not measured) and read the similar factor as a property of the polymer–donor pair rather than of the drug's own rate trasi2014a C9, C12.
  3. PVP raised Tg by only about 2–3 °C, so a change in bulk mobility cannot explain the effect trasi2014a R7; trasi2014b C20.
  4. The inhibition grows as the temperature falls: PVP K-29/32 at 1.5 and 4.5 wt% reduced felodipine growth 2- and 6-fold at 110 °C but 3- and 35-fold at 70 °C. A factor measured at high temperature may therefore understate inhibition in the glass (the extractor's reading) kestur2010b R13; kestur2011 R7; trasi2014a C11.
  5. Some additives do the opposite: TPGS raised celecoxib growth by about two orders at 70 °C, and PHB accelerated growth of the higher-Tg compounds mosqueragiraldo2014 C9; sato2016 R11.
To feel the numbers. At 70 °C neat flutamide grew 78 ÷ 0.47 = about 166 times faster than neat aceclofenac. A tenfold polymer factor turns 78 µm/s into 7.8 µm/s, still 7.8 ÷ 0.47 = about 17 times faster than neat aceclofenac. Reaching the same target rate therefore takes a larger factor, hence more polymer, for the fast grower.
3–14×reduction of melt growth rate by 10 mol% PVP across five drugs, at most temperaturestrasi2014a R6.m, R6.c, X6
≈ 166×neat flutamide against neat aceclofenac at 70 °C, yet both fell by a similar factor with PVPtrasi2014a R6.c, X6
6× → 35×felodipine growth reduction by 4.5 wt% PVP at 110 °C against 70 °Ckestur2010b R13

Connections to other chapters

11 more connections
Evidence: data, conditions and papers

Finding F19. A polymer lowers melt growth rates by a broadly similar factor for fast and slow growers, so fast growers need more polymer

In short. 10 mol% PVP lowered the melt growth rates of five hydrogen-bond-donor drugs by roughly 3- to 14-fold at most temperatures (authors: "around 5−10"), although their neat rates differed widely, about 166-fold between flutamide and aceclofenac at 70 °C (synthesis: 78/0.47) trasi2014a R6.m, R6.c, X6, C10. The authors conclude that faster growers "will clearly require" more polymer for a given rate trasi2014a C11. The factor depended on the polymer, its concentration and the temperature: PVP slowed nilutamide and flutamide growth far more than PAA, in line with stronger hydrogen bonding, and for felodipine, bifonazole and nimesulide the reduction grew at lower temperature and with polymer content trasi2014b R3, C19; trasi2012b C7; kestur2010b R13; kestur2011 R7, R9. Some additives accelerated growth mosqueragiraldo2014 C9; sato2016 R11.

Type. observation in N systems (five drugs with PVP; felodipine, bifonazole, nimesulide, flutamide, acetaminophen, celecoxib, indomethacin and others elsewhere) Strength. measured at several temperatures per drug with consistent ratios for four of five drugs (trasi2014a K5: moderate), but without replicates and on an undefined molar basis (trasi2014a X17, X7).

How it was shown

Trasi and co-workers (2014) chose five drugs with good hydrogen-bond donors, cryomilled each with 10 mol% PVP K-12, melted the mixture between coverslips, seeded it and measured growth at 40–120 °C trasi2014a P3, A2, C9. The basis of "mol%" is not stated trasi2014a X7. Kestur and Taylor (2010) did the same for felodipine with 0.5–4.5% polymer at 70–110 °C kestur2010a P1, P2, P3, P4.

What the data show

The ratios of neat to PVP growth rate were about 9–14 for flutamide, 3–6 for acetaminophen, 4–8 for aceclofenac, 5–8 for indomethacin and about 5 for nilutamide, except about 50 at 70 °C trasi2014a X6. PVP raised Tg by only about 2–3 °C trasi2014a R7.

papersystemmethod and conditionsresultids
trasi2014afive hydrogen-bond-donor drugs10 mol% PVP K-12, 40–120 °C, no replicatesneat/PVP about 9–14 (flutamide), 3–6 (acetaminophen), 4–8 (aceclofenac), 5–8 (indomethacin), 5 (nilutamide)trasi2014a R6, R7, X6, X7, C10
trasi2014bnilutamide10% molar PAA or PVP, 80 °Cabout 5 µm/s alone, 3.8 with PAA, 1.1 with PVPtrasi2014b R3, R11, R12
kestur2010afelodipine form II3% w/w polymers, 70 °Cabout 4-fold (HPMCAS), 7-fold (PVP/VA), 10-fold (PVP) lowerkestur2010a R6
kestur2010bfelodipinePVP K-29/32 at 1.5 and 4.5 wt%2- and 6-fold lower at 110 °C; 3- and 35-fold lower at 70 °Ckestur2010b R13
kestur2011bifonazole; nimesulide5 wt% polymers, 18–100 °Cbifonazole: PAA 5-fold at 100 °C, 15-fold at 18 °C; nimesulide 2–10-foldkestur2011 R7, R9
mosqueragiraldo2014celecoxib10% w/w additivesPVP more than one order lower; TPGS about two orders faster at 70 °Cmosqueragiraldo2014 R1, C9
Fig. 3 of trasi2014a: growth with and without 10 mol% PVP for five drugs
Fig. 3 · trasi2014agrowth with and without 10 mol% PVP for five drugs

Each drug's PVP series runs roughly parallel to and below its neat series on the log axis, for the fast flutamide and the slow indomethacin alike; the one exception is the nilutamide point at 70 °C trasi2014a R6, X6.

Why

The authors attribute growth inhibition mainly to specific drug–polymer interactions: for nilutamide the interaction strength "correlates well" with the inhibition trasi2014b C19. For the five 2014 drugs they assume similar hydrogen bonding with PVP ("would be expected"), which was not measured trasi2014a C9, C12. (synthesis: with a similar factor for each drug, the polymer needed for a target rate scales with the neat rate, which is the authors' conclusion trasi2014a C11.)

Limits and transfer

There are no replicates in the 2014 comparison, and the undefined molar basis means the mass fraction of PVP differs between drugs trasi2014a X17, X7. Only growth was measured, and polymer effects on nucleation can be opposite trasi2012a C46. Measure the neat growth rate and the reduction factor of the candidate polymer at two temperatures, including one close to storage trasi2014a C11; kestur2010b R13.

How it was shown

Kestur and Taylor (2010) diluted 10% w/w polymer–felodipine dispersions with crystalline felodipine in a cryogenic mill to 0.5–4.5% polymer, melt-quenched films between cover glasses, nucleated them at 40 °C and grew form II spherulites at 70–110 °C, in triplicate kestur2010a P1, P2, P3, P4. A companion study did the same with three PVP grades and six PVP concentrations kestur2010b P1, P3, R13. Kestur and Taylor (2011) cryomilled 5% w/w polymers with bifonazole or nimesulide and measured growth from 18 to 100 °C kestur2011 P1, P3. Trasi and co-workers (2012) measured growth of acetaminophen and flutamide with 10% w/w or 10 mol% polymers and additives trasi2012a R6, R8; trasi2012b R3, R5. In 2014 Trasi and co-workers chose five drugs with good hydrogen-bond donors (acetaminophen, flutamide, nilutamide, aceclofenac, indomethacin), cryomilled each with 10 mol% PVP K-12, melted the mixture between coverslips, seeded it with the starting crystal form and measured growth at 40–120 °C trasi2014a P3, A2, C9. The basis of "mol%" (chains or repeat units) is not stated trasi2014a X7. Trasi and Taylor (2014) measured nilutamide growth with 10% molar PAA or with PVP and recorded the drug–polymer hydrogen bonding by FTIR on 1:1 molar films trasi2014b P2, P4. Mosquera-Giraldo and co-workers (2014) and Sato and co-workers (2016) measured celecoxib with surfactants and six compounds with polyhydroxybutyrate (PHB) mosqueragiraldo2014 R1; sato2016 R11.

What the data show

With 10 mol% PVP, read from the 2014 figure, flutamide fell from about 1.5 to 0.14 µm/s at 40 °C and from about 78 to 5.6 µm/s at 70 °C; acetaminophen from about 0.23 to 0.037 µm/s at 60 °C and from about 45 to 7.4 µm/s at 100 °C; aceclofenac from about 0.028 to 0.0035 µm/s at 50 °C; indomethacin from about 1.25 to 0.17 µm/s at 120 °C trasi2014a R6. The ratios were about 9–14 for flutamide, 3–6 for acetaminophen, 4–8 for aceclofenac, 5–8 for indomethacin and about 5 for nilutamide, except about 50 at 70 °C trasi2014a X6. PVP raised Tg by only about 2–3 °C in each mixture trasi2014a R7. The authors conclude that the initial growth rate "did not appear" to influence the extent of reduction trasi2014a C10. For nilutamide at 80 °C, growth was about 5 µm/s alone, 3.8 µm/s with 10% molar PAA and 1.1 µm/s with PVP; PVP shifted the drug NH band from about 3313 to about 3130 cm⁻¹ and its own carbonyl from about 1678 to 1662 cm⁻¹, while PAA left the bands nearly unchanged trasi2014b R3, R12, R11, C17, C18. At 60 °C the PAA point lay above the pure drug trasi2014b X4. Flutamide growth was about one order of magnitude lower with 10 mol% PVP, while PAA had no significant effect trasi2012b R3, C7. For acetaminophen at 10% w/w, PAA lowered growth by more than an order of magnitude, PVP and PVPVA substantially, and HPMCAS, E100 and HPMC much less; at equal mole ratio PAA, PVP and PVPh inhibited similarly trasi2012a C15, C16. With 3% w/w polymer, felodipine growth at 70 °C was about 4-fold lower with HPMCAS, 7-fold with PVP/VA and 10-fold with PVP kestur2010a R6. PVP K-29/32 at 1.5 and 4.5 wt% reduced felodipine growth 2- and 6-fold at 110 °C but 3- and 35-fold at 70 °C kestur2010b R13. For bifonazole, PAA reduced growth 5-fold at 100 °C and 15-fold at 18 °C, PVP and HPMCAS 2–6-fold; nimesulide grew 2–10 times faster without polymer kestur2011 R7, R9. PVP reduced celecoxib growth by more than an order of magnitude, whereas TPGS raised it by about two orders at 70 °C mosqueragiraldo2014 R1, C9. With PHB the growth ratio ran from about 0.42 to 10 across six compounds, acceleration for the higher-Tg compounds sato2016 R11.

papersystemmethod and conditionsresultids
trasi2014aacetaminophen, flutamide, nilutamide, aceclofenac, indomethacin10 mol% PVP K-12 (basis not defined), 40–120 °C, no replicatesneat/PVP ≈9–14 (flutamide), ≈3–6 (acetaminophen), ≈4–8 (aceclofenac), ≈5–8 (indomethacin), ≈5 (nilutamide, ≈50 at 70 °C); Tg raised 2–3 °CR6, R7, X6, X7, C10, C11
trasi2014bnilutamide10% molar PAA or PVP, 80 °C; FTIR of 1:1 films≈5 (alone), ≈3.8 (PAA), ≈1.1 µm/s (PVP); PVP shifts the drug NH and its own C=O bandsR3, R11, R12, C19
kestur2010afelodipine form II3% w/w polymers, 70 °C≈4-fold (HPMCAS), ≈7-fold (PVP/VA), ≈10-fold (PVP) lowerR6
kestur2010bfelodipinePVP K-29/32 at 1.5 and 4.5 wt%2- and 6-fold lower at 110 °C; 3- and 35-fold lower at 70 °CR13
kestur2011bifonazole; nimesulide5 wt% polymers, 18–100 °Cbifonazole: PAA 5-fold at 100 °C, 15-fold at 18 °C; nimesulide 2–10-foldR7, R9
trasi2012bflutamide10 mol% additivesPVP about one order lower; PAA not significant; hydrogen bonding acts on growth but not on nucleationR3, C7, C45
trasi2012aacetaminophen10% w/w polymersPAA more than one order lower growth but more nucleation; HPMCAS the best nucleation inhibitor and a weak growth inhibitorR6, C15, C23, C46
mosqueragiraldo2014celecoxib10% w/w additivesPVP more than one order lower; TPGS about two orders faster at 70 °CR1, C9
sato2016six compoundsPHBratio of growth with to without PHB ≈0.42 to ≈10, acceleration for the higher-Tg compoundsR11

Figure

Fig. 3 of trasi2014a: growth with and without 10 mol% PVP for five drugs
Fig. 3 · trasi2014agrowth with and without 10 mol% PVP for five drugs

Each drug's PVP series runs roughly parallel to and below its neat series on the log axis, for the fast flutamide and the slow indomethacin alike; the one exception is the nilutamide point at 70 °C trasi2014a R6, X6.

Fig. 2 of trasi2014b: stable-form nilutamide growth, pure and with PAA or PVP
Fig. 2 · trasi2014bstable-form nilutamide growth, pure and with PAA or PVP

Blue (nilutamide), green (PAA) and red (PVP) on a log scale: PAA lies just below the pure drug, PVP far below, most of all at the lowest temperature trasi2014b R3.

Why

The authors attribute growth inhibition mainly to specific drug–polymer interactions: the interaction strength "correlates well" with the inhibition for nilutamide, PVP being a better hydrogen-bond acceptor than the drug itself ("this is possible because") trasi2014b C19, C18. For the five 2014 drugs, all with good donors, they assume similar hydrogen bonding with PVP ("would be expected"), which was not measured, and read the similar factor as a property of the polymer–donor pair rather than of the drug's intrinsic rate trasi2014a C9, C12. Hydrogen bonding has "a very clear effect" on growth but not on nucleation trasi2012b C45. The low polymer levels changed Tg by only a few degrees, so a change in bulk mobility as measured by Tg cannot explain the effect trasi2014a R7; trasi2014b C20. For felodipine, the PVP effect "is believed to be kinetic rather than thermodynamic", since PVP hardly lowers the driving force at these levels kestur2010b C28. The growth-rate ratios of felodipine forms I and II with PVP were similar, which the authors read as PVP acting on the liquid phase rather than at the crystal surface ("most likely") kestur2013a C6. Stronger inhibition at low temperature is attributed "most likely" to reduced mobility and a smaller driving force together tian2017 C31. Accelerators such as TPGS and PHB act the other way, PHB "most likely" through the mobility of the solute sato2016 C19; mosqueragiraldo2014 C9. (synthesis: if a polymer divides the growth rate by a similar factor for each drug, the polymer needed to reach a target rate scales with the neat rate, which is the authors' conclusion; the felodipine and bifonazole data add that the factor itself grows as the temperature falls toward storage conditions, so a factor measured at high temperature may understate inhibition in the glass trasi2014a C11; kestur2010b R13; kestur2011 R7.)

Conditions and limits

Neat melts with low polymer levels, 1.5–10% by weight or 10 mol%, above Tg trasi2014a P3; kestur2010b R13; kestur2011 P1. No replicates in the 2014 comparison, and the undefined molar basis means the mass fraction of PVP differs between drugs of molecular weight 151.1 to 357.7 trasi2014a X17, X7. Only growth was measured; polymer effects on nucleation were studied separately and can be opposite (see F3) trasi2012a C46. Drugs without hydrogen-bond donors were not tested trasi2014a C12.

Counterexamples and conflicts

Nilutamide was reduced about 50-fold at 70 °C trasi2014a X6. Inhibition factors rose strongly at low temperature for felodipine and bifonazole kestur2010b R13; kestur2011 R7. Some additives accelerated growth mosqueragiraldo2014 C9; sato2016 R11. In solution, a fast-growing drug (efavirenz) was inhibited less by polymers than slower ones, which the authors read as fast growers being harder to retard ilevbare2013b C5.

Transfer check

Measure the neat growth rate and the reduction factor of the candidate polymer at two temperatures, including one close to storage, and choose the loading for the rate needed trasi2014a C11; kestur2010b R13. Check that the drug has a hydrogen-bond donor and the polymer a good acceptor, by FTIR on drug–polymer films, before expecting a PVP-like factor trasi2014a C12; trasi2014b C18. Test growth and nucleation separately, because the best growth inhibitor can promote nucleation trasi2012a C46.

History

Kestur in 2010 and 2011 showed concentration- and temperature-dependent growth inhibition kestur2010a R6; kestur2010b R13; kestur2011 R7. Trasi in 2012 separated polymer effects on nucleation and growth trasi2012a C46; trasi2012b C45. Trasi in 2014 compared the factor across drugs of very different growth rates and tied it to hydrogen bonding in nilutamide trasi2014a C10; trasi2014b C19. Accelerators were reported in 2014 and 2016 mosqueragiraldo2014 C9; sato2016 R11.

F20

Topic 4Formulation consequences3 of 4

Making a salt raised the glass transition for every counterion tested, but crystallization and moisture uptake did not follow acid strength.

Every amorphous salt of propranolol and nicardipine had a higher Tg than its free base, and strong-acid counterions such as chloride gave the highest values. But the highest-Tg propranolol salt crystallized on heating and at 79% RH, and moisture uptake at 75% RH ranged from 0.39 to 61.90% with no link to the counterion pKa (a measure of acid strength) towler2008 R4, R5, C7, C8, R12, R8, C14.

moderateStrong for the Tg rise, which is large and measured directly; weak to moderate for crystallization and moisture, which rest on a few salts per stress with unreported stoichiometry towler2008 K1, K6, K5, X8.

free base (± spread)range of the amorphous saltspropranololacetate to hydrochloridebase −9.3salts 9.0–35.9nicardipinebenzoate to hydrochloridebase 3.2salts 28.5–81.0−20−100102030405060708090glass transition temperature, °C

Every salt lies above its free base. towler2008 R4, R5

baserange of the saltspropranololcrystalline base 2.54salts 0.39–61.90nicardipineamorphous base 2.890salts 4.2–12.40.1110100moisture uptake at 75% RH, % of weight (log scale)

Tosylate was the lowest and acetate the highest propranolol salt; the authors found no apparent link to pKa. towler2008 R8, R9, C14

For your drug

  1. Screen counterions for Tg by DSC first, with a cycling protocol that dries the sample.
  2. Then test crystallization separately under heat and under humidity (for example 40 °C/75% RH or 79% RH storage with PXRD or microscopy) and measure moisture sorption, because neither followed Tg or pKa reliably.
  3. Report the base:acid ratio and confirm salt formation spectroscopically. If the salt goes into a polymer dispersion, test the dispersion in the dissolution medium too, because salt formation may not change crystallization there.

towler2008 P6, P9, C33, C14, X8; hiew2022b R7; duong2022a C47; nguyen2023b C17

How it works and key numbers
  1. From polymer work: stronger intermolecular interactions need more thermal energy for mobility and so raise Tg towler2008 C21.
  2. For salts, the authors argue that the ionic interaction between the protonated base and the counterion "will be of greater importance" than hydrogen bonding, and they "believe" the salt Tg is strongly affected by this acid–base interaction. The interaction itself was not measured towler2008 C23, C25.
  3. "For the majority" of salts Tg fell as counterion pKa rose, but propranolol tosylate and phosphate sat below salts of higher pKa; counterion size and shape are "likely" to explain why the ranking is not linear towler2008 C8, X14, C9, C26.
  4. For crystallization and moisture uptake the authors offer no mechanism: "more work needs to be performed" towler2008 C34.
  5. The extractor's reading: Tg rose with counterion strength in both series, but crystallization depended on the stress and the salt, so Tg is a necessary but not sufficient guide. Propranolol hydrochloride had the highest Tg yet crystallized, while lumefantrine hydrochloride was among the most stable salts towler2008 R4.a, R12; hiew2022b R7.
  6. Later work: delamanid sulfonate salts (Tg 83.7–117.8 °C against 43.3 °C for the free base) stayed amorphous for 2 weeks at 40 °C/75% RH, and the crystallization order of lumefantrine salts broadly followed Tg duong2022a R20, M1, C32; hiew2022b R2, R7.
To feel the numbers. Hydrochloride against free base: propranolol 35.9 − (−9.3) = 45.2 °C higher; nicardipine 81.0 − 3.2 = 77.8 °C higher.
at least +18.3 °C and +25.3 °Csmallest Tg rise over the free base, propranolol and nicardipinetowler2008 R4.j, R4.k, R5.h, R5.i
0.39 → 61.90%moisture uptake of propranolol salts at 75% RH, with no pKa linktowler2008 R8, C14
11 of 20salts amorphous after freeze-drying (hydrochloride, phosphate, tartrate always; benzoate, acetate never)towler2008 R1, C6

Connections to other chapters

Evidence: data, conditions and papers

Finding F20. Salt formation raises the Tg of an amorphous base for every counterion tested, but crystallization and moisture uptake do not follow counterion pKa

In short. Every amorphous salt of propranolol and nicardipine had a higher Tg than its free base, and strong-acid counterions such as chloride gave the highest values, with exceptions towler2008 R4, R5, C7, C8, X14. Crystallization under freeze-drying, heating and humidity changed with the counterion but did not follow Tg or pKa: the highest-Tg propranolol salt crystallized on heating and at 79% RH, and moisture uptake at 75% RH spanned 0.39–61.90% without a pKa link towler2008 R4, R12, R8, C14, C34. Later library papers used salt formation to raise the Tg and stability of amorphous delamanid and lumefantrine, the lower-Tg salts being the least stable duong2022a C47; hiew2022b R7, C37.

Type. observation in N systems (two amine bases with 19 identifiable salts; delamanid and lumefantrine salts later) Strength. for the Tg rise, which is large and measured directly (towler2008 K1: strong); weak to moderate for crystallization and moisture, which rest on a few salts per stress with unreported stoichiometry (towler2008 K6, K5: moderate; section 0.8: moderate).

How it was shown

Towler and co-workers (2008) chose propranolol (pKa 9.5) and nicardipine (pKa 8.6) so that many acids could protonate them towler2008 M1, M2, C5. Each free base was mixed with an aqueous acid and freeze-dried; salts not amorphous afterwards were melt-quenched towler2008 P1, P2, P3, P5. Tg was taken by DSC from the last of several cycles, which dried the sample towler2008 P6. Moisture uptake was measured by vapour sorption, and samples were stored at 0% and then 79% RH towler2008 P8, P9.

What the data show

Salt Tg lay between 9.0 and 35.9 °C for propranolol and between 28.5 and 81.0 °C for nicardipine; "for the majority" of salts Tg fell as counterion pKa rose towler2008 R4, R5, C8. Crystallization behaved differently from Tg towler2008 C33, C34.

papersystemmethod and conditionsresultids
towler2008propranolol and its saltsfreeze-dried or melt-quenched; DSC at 20 °C/minbase −9.3 °C; salts 9.0 (acetate) to 35.9 °C (hydrochloride)towler2008 R4, C7
towler2008nicardipine and 8 saltsas abovebase 3.2 °C; salts 28.5 (benzoate) to 81.0 °C (hydrochloride)towler2008 R5, C7
towler2008crystallizationfreeze-drying; DSC heating; storage at 79% RH11 of 20 amorphous after freeze-drying; four propranolol salts recrystallized on heating (46.5–105.4 °C), none for nicardipine; hydrochloride and tosylate crystallized at 79% RHtowler2008 R1, R4, R5, R12
towler2008moisturevapour sorption, 75% RHpropranolol salts 0.39–61.90%, nicardipine salts 4.2–12.4%; no apparent pKa linktowler2008 R8, R9, C14
duong2022adelamanid and 7 sulfonate saltsopen dish, 40 °C/75% RH, 2 weekssalts Tg 83.7–117.8 °C against 43.3 °C; all stayed amorphousduong2022a R20, M1, C32
hiew2022bneat amorphous lumefantrine saltsopen dish, 40 °C/75% RH, PXRDbenzoate, PEG 250 diacid and besylate crystallized within 4 weeks; camsylate after 32 weeks; HCl and sulfate not for at least 16 weekshiew2022b R2, R7, C37
Fig. 5 of towler2008: propranolol tosylate before and after storage at 79% RH
Fig. 5 · towler2008propranolol tosylate before and after storage at 79% RH

Irregular amorphous particles (a) have turned into needle and plate crystals (b), so a raised Tg did not prevent moisture-induced crystallization towler2008 R13.

Why

The authors take from polymer work that stronger intermolecular interactions raise Tg towler2008 C21, and argue that ionic interactions between the protonated base and the counterion "will be of greater importance" than hydrogen bonding towler2008 C23. The interaction itself was not measured towler2008 C25. For crystallization and moisture uptake they offer no mechanism: "more work needs to be performed" towler2008 C34. (synthesis: Tg is a necessary but not sufficient guide towler2008 R4.a, R12; hiew2022b R7.)

Limits and transfer

Only two amine bases were studied towler2008 A2. Salt stoichiometry was not reported, and several standard deviations exceed the differences between neighbouring salts towler2008 X8, X17. Screen counterions for Tg first, then test crystallization under heat and humidity separately and measure moisture sorption towler2008 P6, P9, C33, C14.

How it was shown

Towler and co-workers (2008) asked whether the counterion changes the Tg, crystallization tendency and moisture uptake of an amorphous salt, and which counterion properties go with a high Tg towler2008 A1. Propranolol (secondary amine, pKa 9.5, MW 259.3) and nicardipine (tertiary amine, pKa 8.6, MW 479.5) were chosen so that many acids could protonate them towler2008 M1, M2, C5. Each free base was precipitated from its hydrochloride with NaOH, dissolved at 20 mg/mL (propranolol in ethanol, nicardipine in acetone), mixed with an aqueous acid at a molar ratio that is called "required" but not printed, and freeze-dried (frozen at −45 °C, dried at 100 mT with the shelf at −30 and −10 °C, then at 5 °C) towler2008 P1, P2, P3, X8. Salts not amorphous after freeze-drying were melt-quenched, under conditions not printed towler2008 P5. Up to ten counterions per base spanned counterion pKa −6 (hydrochloride) to 4.756 (acetate) towler2008 A1, R4, R5. Tg was taken by DSC (about 8 mg, pinholed pans, 20 °C/min) from the last of several cycles between −40 °C and about 20 °C above Tg, which erased thermal history and dried the sample; a screening scan to 250 °C recorded recrystallization (Tc) and melting towler2008 P6. Moisture uptake was measured by vapour sorption from 5 to 95% RH in 10% steps, and samples were stored at 0% and then 79% RH and inspected by polarized light microscopy (5 days in the text, 48 h in the captions) towler2008 P8, P9, X4. A PLS2 model related salt Tg to counterion descriptors, including DFT electronic descriptors of the isolated acids towler2008 P10, P11. The n behind each ± and the Tg convention are not stated towler2008 X7.

What the data show

Propranolol free base had Tg −9.3 ± 4.0 °C, and its nine salts with a Tg lay between 9.0 ± 5.6 (acetate) and 35.9 ± 0.6 °C (hydrochloride) towler2008 R4. Nicardipine free base had Tg 3.2 ± 9.6 °C, and its eight salts lay between 28.5 ± 3.1 (benzoate) and 81.0 ± 4.2 °C (hydrochloride) towler2008 R5. The rise over the free base was therefore at least 18.3 °C for propranolol and 25.3 °C for nicardipine (synthesis: 9.0 − (−9.3) and 28.5 − 3.2) towler2008 R4.j, R4.k, R5.h, R5.i. "For the majority" of salts Tg fell as counterion pKa rose, but propranolol tosylate (pKa −1.34, 16.7 °C) and phosphate (1.96, 11.3 °C) sat below salts of higher pKa, and nicardipine besylate (0.7, 65.8 °C) lay below the oxalate (1.271, 71.3 °C) towler2008 C8, X14. Crystallization behaved differently from Tg towler2008 C33, C34. Of 20 salts prepared, 11 were amorphous after freeze-drying: hydrochloride, phosphate and tartrate always, benzoate and acetate never towler2008 R1, C6. On DSC heating four propranolol salts recrystallized, the hydrochloride at 105.4, besylate 46.5, tartrate 80.1 and acetate 78.4 °C, against 31.2 °C for the free base; no nicardipine salt recrystallized towler2008 R4, R5, C10. After storage at 79% RH, propranolol hydrochloride and tosylate had crystallized into needles and plates, while the tartrate showed no crystals and looked deliquesced towler2008 R12, R13, R14, C17. During sorption propranolol hydrochloride, tosylate and tartrate gained and then lost weight, read as crystallization; no nicardipine salt did towler2008 R11, C15. At 75% RH propranolol salts gained 0.39% (tosylate) to 61.90% (acetate) and nicardipine salts 4.2% (oxalate) to 12.4% (besylate), against 2.54% for the crystalline propranolol base and 2.890% for the amorphous nicardipine base towler2008 R8, R9. The authors found "no apparent link" between moisture uptake and counterion pKa towler2008 C14. A PLS2 model on 12 salts, after citrate and phosphate were removed as outliers, gave R² 0.909, Q² 0.668 and RMSEE 3.8 and 3.4 °C, with pKa weighted negatively for both drugs towler2008 R16, R17, R20, C29. In later work, seven sulfonate salts of delamanid were amorphous with Tg 83.7–117.8 °C against 43.3 °C for the free base and stayed amorphous for 2 weeks at 40 °C/75% RH duong2022a R20, M1, C32. Neat amorphous lumefantrine salts (free base Tg 19.7 °C, salts up to about 101 °C for the sulfate) crystallized at 40 °C/75% RH in an order that broadly followed Tg: benzoate, PEG 250 diacid and besylate within 4 weeks, camsylate after 32 weeks, hydrochloride and sulfate not for at least 16 weeks hiew2022b R2, R7.

papersystemmethod and conditionsresultids
towler2008propranolol and 10 saltsfreeze-dried or melt-quenched; DSC, final heating at 20 °C/minbase −9.3 °C; salts 9.0 (acetate) to 35.9 °C (hydrochloride); no Tg for the besylateR4, R2, C7
towler2008nicardipine and 8 saltsas abovebase 3.2 °C; salts 28.5 (benzoate) to 81.0 °C (hydrochloride)R5, C7
towler2008both basesTg against counterion pKa −6 to 4.756falls with pKa "for the majority"; tosylate, phosphate and nicardipine besylate break the orderC8, C9, X14
towler2008crystallizationfreeze-drying; DSC heating; storage at 79% RH11 of 20 amorphous after freeze-drying; Tc 46.5–105.4 °C for four propranolol salts, none for nicardipine; hydrochloride and tosylate crystallized at 79% RHR1, R4, R5, R12, C32, C33
towler2008moisturevapour sorption, 75% RHpropranolol salts 0.39–61.90%, nicardipine salts 4.2–12.4%; no apparent pKa linkR8, R9, C14
towler2008PLS2 model7 propranolol and 5 nicardipine saltsR² 0.909, Q² 0.668, RMSEE 3.8 and 3.4 °CR16, R17, X11
duong2022adelamanid and 7 sulfonate saltsopen dish, 40 °C/75% RH, 2 weekssalts Tg 83.7–117.8 °C against 43.3 °C; all stayed amorphous, four liquefiedR20, M1, C32, C47
hiew2022bneat amorphous lumefantrine saltsopen dish, 40 °C/75% RH, PXRDbenzoate, PEG 250 diacid and besylate crystallized within 4 weeks; camsylate after 32 weeks; HCl and sulfate not for at least 16 weeksR2, R7, C37
nguyen2023bdelamanid free base and salts in dispersion filmspH 3.0 buffersalt formation did not change the crystallization tendencyC17

Figure

Fig. 2 of towler2008: Tg against counterion pKa for propranolol salts, free-base Tg as a dotted line
Fig. 2 · towler2008Tg against counterion pKa for propranolol salts, free-base Tg as a dotted line

Every point lies above the dotted free-base line near −9 °C, and the trend falls with pKa; tosylate (pKa −1.34) and phosphate (1.96) sit below it towler2008 R6, X14.

Fig. 5 of towler2008: propranolol tosylate before and after storage at 79% RH
Fig. 5 · towler2008propranolol tosylate before and after storage at 79% RH

Irregular amorphous particles (a) have turned into needle and plate crystals (b), so a raised Tg did not prevent moisture-induced crystallization towler2008 R13.

Why

The authors take from polymer work that stronger intermolecular interactions need more thermal energy for mobility and so raise Tg towler2008 C21. For salts, they argue, ionic interactions between the protonated base and the counterion "will be of greater importance" than hydrogen bonding, as the earlier indomethacin salt work had shown towler2008 C23, C3. They "believe" that the Tg of the amorphous salts is strongly affected by the acid–base interaction, and they read the negative PLS weights of pKa and electronic chemical potential, and the positive weight of the electrophilicity index for propranolol, as descriptors that "can be related to" that interaction strength towler2008 C25, C31, R20. The interaction itself was not measured, and the descriptors were calculated for single acid molecules towler2008 C25, P10. Counterion size and shape are "likely" to explain why the pKa ranking is not linear, and citrate and phosphate are outliers "probably" because of their trivalent character, although salt stoichiometry was not reported towler2008 C9, C26, X8. For crystallization and moisture uptake the authors offer no mechanism: "more work needs to be performed" towler2008 C34. Hiew and co-workers later read the drug or salt Tg as "an important predictor" of dispersion stability, especially at higher drug loading hiew2022b C37. (synthesis: in both 2008 series Tg rose with counterion strength, but crystallization depended on the stress and the salt, so Tg is a necessary but not sufficient guide; the propranolol hydrochloride, highest in Tg yet crystallizing under heat and humidity, and the lumefantrine hydrochloride, among the most stable salts, show the same counterion behaving differently with different bases towler2008 R4.a, R12; hiew2022b R7.)

Conditions and limits

Two amine bases, freeze-dried or melt-quenched neat salts, tested at 0% and 79% RH for 48 h or 5 days and in DSC heating at 20 °C/min towler2008 P3, P5, P6, P9, X4. Salt stoichiometry and the degree of ionization in the amorphous solid were not reported towler2008 X8. Several sorption samples were not amorphous after freeze-drying, and their preparation for sorption is not stated towler2008 X6. Several standard deviations are larger than the differences between neighbouring salts, so close salts cannot be ranked towler2008 X17. The tartrate's crystallization under humidity, listed in the conclusions, is not confirmed by the storage test towler2008 X5. The PLS model rests on 12 salts with about 17 descriptors and was not tested on new salts towler2008 X11, X12. Only two amine bases were studied in 2008, and no polymer dispersion of a salt towler2008 A2, A1.

Counterexamples and conflicts

The high-Tg propranolol hydrochloride crystallized on heating and at 79% RH towler2008 R4.a, R12, whereas the lumefantrine hydrochloride did not recrystallize for at least 16 weeks at 40 °C/75% RH hiew2022b R7. In delamanid dispersion films in pH 3.0 buffer, salt formation did not change the crystallization tendency nguyen2023b C17. The text names the acetate as the lowest-Tg salt for both drugs, but no nicardipine acetate Tg exists; the lowest nicardipine values are the benzoate and citrate towler2008 X2. The stated Tc ranking with pKa is broken by the besylate, whose Tc of 46.5 °C is the lowest towler2008 X3.

Transfer check

Screen counterions for Tg by DSC first, with a cycling protocol that dries the sample towler2008 P6. Then test crystallization separately under heat and under humidity (for example 40 °C/75% RH or 79% RH storage with PXRD or microscopy), and measure moisture sorption, because neither followed Tg or pKa reliably in the 2008 data towler2008 P9, C33, C14; hiew2022b R7. Report the base:acid ratio and confirm salt formation spectroscopically, since stoichiometry was the main unreported variable in 2008 towler2008 X8; duong2022a C47. If the salt goes into a polymer dispersion, test the dispersion in the dissolution medium too, because salt formation may not change crystallization there nguyen2023b C17.

History

Counterion effects on the Tg of amorphous indomethacin salts were reported in 2002 and are cited towler2008 C3. Towler in 2008 extended them to organic counterions and two amine bases, finding a broad pKa trend in Tg but no rule for crystallization or moisture towler2008 C35, C34. Duong in 2022 used sulfonate salts to raise the Tg of delamanid and stabilize it against crystallization duong2022a C47, C66. Hiew in 2022 found salt Tg to be "an important predictor" of lumefantrine dispersion stability hiew2022b C37. Nguyen in 2023 found no salt effect on crystallization of delamanid dispersion films at pH 3.0 nguyen2023b C17.

F21

Topic 4Formulation consequences4 of 4

Crystals form and grow faster on a free surface than in the confined bulk, so sandwiched-film screens can understate the risk.

Where both were measured, a free surface crystallized faster than the confined bulk: felodipine form I grew about 6 times faster at the surface at 30 °C, and nilutamide nucleated much faster at the surface kestur2013b R12; kestur2013a R9; trasi2014b R1. Screens that store glasses between two cover glasses may therefore understate the crystallization of powders and open surfaces baird2010 C45.

moderateModerate: direct surface and bulk measurements in two drugs and a 14-drug comparison, but the 14-drug surface and bulk data lie at different temperatures trasi2014a X16; kestur2013a R8.

Felodipine form I growth at 30 °C, 10⁻⁹ m/sfree surface2.7bulk0.47

A ratio of 5.7; with 3% PVP the ratio rose to 22.8. kestur2013b R12

surface faster than bulkabout 6 times at 18 °Cbelow 65 °Cno differenceabove 70 °C182838485868788898temperature, °C

Felodipine form I, free surface against bulk. kestur2013a R9

For your drug

  1. Test open-surface films or powders as well as sandwiched films, at storage-relevant temperatures near or below Tg, where the surface advantage appears.
  2. Inspect particle and tablet surfaces during stability and dissolution studies, for example by SEM, which revealed surface-grown crystals in these studies.
  3. For high-load spray-dried dispersions that crystallize at the surface within weeks, a surface barrier such as an alumina coating is the strategy the library has tested.

kestur2013a R9; kestur2013b R12; duong2022c R16, C33, C35; correasoto2017 C26

How it works and key numbers
  1. Surface growth is "generally" faster than bulk growth, especially at low temperature where the material does not flow, and "is thought to" reflect higher molecular mobility at the surface near or below Tg trasi2014a C4.
  2. For felodipine, growth jumps below 50 °C for both surface and bulk crystals, "most likely" a glass–crystal growth mode activated near Tg. Extrapolating high-temperature rates to storage "may not be predictive" and may give "an over prediction of the stability" kestur2013a C3, C7, C18.
  3. The 2010 storage test and the 2014 bulk growth rates used sandwiched geometries, so their rankings refer to bulk behaviour (the extractor's reading) baird2010 C45; trasi2014a P1.
  4. Surface and bulk growth also rank drugs differently: felodipine is fast at the surface but slow in the bulk trasi2014a R4, R5, C8.
  5. In spray-dried dispersions, crystals grow as needles out of the particle surfaces. In ezetimibe dispersions stored open at 40 °C/75% RH, crystals appeared on uncoated particles after 19 days at 50% drug loading and after 12 days at 70%, while coated particles showed no detectable crystals for two years duong2022c R16, R17, C33; correasoto2017 R16, C26.
Stability of spray-dried dispersions, monthsuncoatedwithin 1 month5–40 nm alumina coatingstable 6–24 months

Dispersions that crystallized within a month uncoated, across earlier coating studies; the coated bar is drawn at the top of the 6–24 month range. moseson2023 R27

2.7 vs 0.47 × 10⁻⁹ m/sfelodipine form I growth at the surface against the bulk, 30 °Ckestur2013b R12
below 65 °Ctemperatures where felodipine surface growth exceeds bulk growth, about 6 times at 18 °Ckestur2013a R9
19 and 12 daysezetimibe dispersions: first crystals on uncoated particles at 50% and 70% loadingduong2022c R16, R17

Connections to other chapters

2 more connections
Evidence: data, conditions and papers

Finding F21. Free surfaces crystallize faster than confined bulk, so screens in sandwiched films can understate the risk

In short. Where both were measured, crystallization at a free surface outpaced the bulk: felodipine form I grew about 6 times faster at the surface than in the bulk at 30 °C, the gap opening only below about 65 °C, and nilutamide nucleated much faster at the surface kestur2013b R12; kestur2013a R9; trasi2014b R1. Surface and bulk growth also ranked drugs differently, felodipine being fast at the surface but slow in the bulk trasi2014a R4, R5, C8. The storage test of the 2010 screen used glasses between two cover glasses, assumed to suppress surface crystallization, so such screens may understate the crystallization of powders and open surfaces (synthesis: from the cover-glass assumption and the surface data) baird2010 C45. Coating particle surfaces delayed crystallization of high-load dispersions from weeks to many months duong2022c C33, C35; moseson2023 R27.

Type. observation in N systems (felodipine, nilutamide, 14 drugs surface against bulk; coated dispersions) Strength. direct surface and bulk measurements in two drugs and a 14-drug comparison, but the 14-drug surface and bulk data lie at different temperatures (trasi2014a X16).

How it was shown

Kestur and Taylor (2013) measured growth of felodipine form I at the free surface and in the bulk from 18 to 100 °C, three samples per point, and at 30 °C with and without 3% PVP kestur2013a R8, R9; kestur2013b R12. Trasi and co-workers (2014) compared surface and bulk growth of 14 drugs trasi2014a P2, R4, R5. Later papers stored spray-dried dispersions uncoated or coated with alumina for up to two years duong2022c R16, R17, R18.

What the data show

Surface and bulk rates did not differ above 70 °C, and the surface was faster below 65 °C kestur2013a R9. In ezetimibe dispersions stored open at 40 °C/75% RH, crystals appeared on uncoated particles after 19 days at 50% drug loading and after 12 days at 70% duong2022c R16, R17.

papersystemmethod and conditionsresultids
kestur2013bfelodipine form I30 °C, 0% RH2.7 × 10⁻⁹ (surface) against 0.47 × 10⁻⁹ m/s (bulk), ratio 5.7; with 3% PVP ratio 22.8kestur2013b R12
kestur2013afelodipine form I18–100 °C, n = 3no difference above 70 °C; surface faster below 65 °C; about 6 times at 18 °Ckestur2013a R8, R9
trasi2014a14 drugssurface growth near Tg (T/Tg about 0.98–1.14) against bulk (about 1.07–1.33)surface rates about 0.00005 (loratadine) to about 1 µm/s (nimesulide); felodipine fast at the surface yet slow in the bulktrasi2014a R4, R5, C8, X16
duong2022cezetimibe spray-dried dispersions40 °C/75% RH, open dish, SEM and PXRD, two yearsuncoated particles carried crystals after 19 days (50% loading) and 12 days (70%); coated mostly crystal-free for two yearsduong2022c R16, R17, R18, C33, C35
baird2010stored glassesbetween two cover glassesassumed to minimize surface crystallization; not testedbaird2010 C45
Fig. 2 of kestur2013a: felodipine growth rate against temperature at the free surface and in the bulk
Fig. 2 · kestur2013afelodipine growth rate against temperature at the free surface and in the bulk

Follow the two form I series: they coincide above about 70 °C and separate below 65 °C, the surface series staying above the bulk down to 18 °C kestur2013a R8, R9.

Why

Surface growth is "generally" faster than bulk growth, and "is thought to" reflect higher molecular mobility at the surface near or below Tg trasi2014a C4. Where such faster modes occur, extrapolating high-temperature growth rates to storage temperatures "may not be predictive" kestur2013a C7, C18. (synthesis: the 2010 storage test and the 2014 bulk growth rates used sandwiched geometries, so their rankings refer to bulk behaviour baird2010 C45; trasi2014a P1.)

Limits and transfer

Surface and bulk growth were measured in different T/Tg windows in the 14-drug comparison trasi2014a X16. The felodipine surface–bulk data are for one polymorph of one drug kestur2013a R8. Test open-surface films or powders as well as sandwiched films, at storage-relevant temperatures near or below Tg kestur2013a R9; kestur2013b R12.

How it was shown

Baird and co-workers (2010) stored quenched glasses between two microscope cover glasses at 0% RH and assumed, without testing, that this minimized surface crystallization baird2010 P7, C45. Kestur and Taylor (2013) measured growth of felodipine form I at the free surface and in the bulk from 18 to 100 °C, three samples per point kestur2013a R8, R9. In a second 2013 study they measured both at 30 °C and 0% RH, with and without 3% PVP, alongside the crystallization of powders of two particle sizes kestur2013b R12, R3. Trasi and co-workers (2014) measured surface growth of 14 drugs near Tg by removing the top coverslip, and compared it with bulk growth between coverslips trasi2014a P2, R4, R5. Trasi and Taylor (2014) counted nilutamide nuclei on open glass surfaces because nucleation in the bulk was too slow trasi2014b P7, R1. Later ASD-process papers stored spray-dried high-load dispersions uncoated or coated with nanometre aluminium oxide layers at 40 °C and high humidity and followed surface crystals by SEM and PXRD for up to two years duong2022c R16, R17, R18; moseson2022a R11.

What the data show

Felodipine form I grew at 2.7 × 10⁻⁹ m/s at the surface against 0.47 × 10⁻⁹ m/s in the bulk at 30 °C, a ratio of 5.7; with 3% PVP the ratio rose to 22.8 kestur2013b R12. Over 18–100 °C, surface and bulk rates did not differ above 70 °C, the surface was faster below 65 °C, and it was about 6 times faster at 18 °C kestur2013a R9. Nilutamide's bulk nucleation rate was very low and its surface rate much higher, with no numbers printed trasi2014b R1. Among 14 drugs, surface rates near Tg ran from about 0.00005 µm/s (loratadine) to about 1 µm/s (nimesulide) trasi2014a R4. Felodipine grew faster at the surface (about 0.06–0.42 µm/s at T/Tg 1.04–1.11) than carbamazepine and nilutamide, yet in the bulk it was slower than nilutamide at all temperatures and than carbamazepine at higher temperatures trasi2014a R4, R5, C8. Loratadine was slowest in both modes, and piroxicam and acetaminophen fast in both trasi2014a C8. The surface data lie at T/Tg of about 0.98–1.14 and the bulk data at about 1.07–1.33 trasi2014a X16. In spray-dried ezetimibe dispersions stored open at 40 °C/75% RH, crystals appeared on uncoated particles after 19 days at 50% drug loading and after 12 days at 70%, while coated particles showed no detectable crystals for two years, apart from a few surface crystals after 7 months on the thinnest coating at 70% duong2022c R16, R17, C33. The drug crystallized more slowly with lower loading and thicker coating duong2022c C35. Across earlier coating studies, ASDs that crystallized within a month uncoated were stable for 6–24 months with 5–40 nm alumina coatings moseson2023 R27. In other dispersions, crystals grew as needles out of spray-dried particle surfaces or formed on tablet surfaces during dissolution correasoto2017 R16, C26; saboo2020 R9, C17.

papersystemmethod and conditionsresultids
kestur2013bfelodipine form I30 °C, 0% RH2.7 × 10⁻⁹ (surface) against 0.47 × 10⁻⁹ m/s (bulk), ratio 5.7; with 3% PVP ratio 22.8R12
kestur2013afelodipine form I18–100 °C, n = 3no difference above 70 °C; surface faster below 65 °C; about 6 times at 18 °CR8, R9, C18
trasi2014bnilutamidemelt-cooled filmsvery low nucleation in the bulk, much higher at the surfaceR1
trasi2014a14 drugssurface growth near Tg (T/Tg ≈0.98–1.14) against bulk (≈1.07–1.33)surface rates ≈0.00005 (loratadine) to ≈1 µm/s (nimesulide); felodipine ≈0.06–0.42 µm/s at the surface yet slow in the bulkR4, R5, C8, X16
baird2010stored glassesbetween two cover glassesassumed to minimize surface crystallization; not testedC45
duong2022cezetimibe spray-dried dispersions40 °C/75% RH, open dish, SEM and PXRD, two yearsuncoated particles carried crystals after 19 days (50% loading) and 12 days (70%); coated mostly crystal-free for two yearsR16, R17, R18, C33, C35
moseson2023Al2O3-coated dispersions (cited earlier studies)open dish, highly accelerated storagecoatings extended stability from below 1 month to 6–24 monthsR27, C1
correasoto2017FTM–HPMCAS spray-dried dispersion40 °C, 75 or 82% RH, 60 days, SEMneedle-shaped crystals growing outward from particle surfacesR16, C26

Figure

Fig. 2 of kestur2013a: felodipine growth rate against temperature at the free surface and in the bulk
Fig. 2 · kestur2013afelodipine growth rate against temperature at the free surface and in the bulk

Follow the two form I series: they coincide above about 70 °C and separate below 65 °C, the surface series staying above the bulk down to 18 °C, below Tg kestur2013a R8, R9.

Fig. 2 of trasi2014a: surface against bulk growth for 14 drugs
Fig. 2 · trasi2014asurface against bulk growth for 14 drugs

Panel (a), surface rates near Tg, ranks the drugs differently from panel (b), bulk rates at higher T/Tg; note that the two panels cover different T/Tg windows trasi2014a R4, R5, X16.

Why

Surface growth is "generally" faster than bulk growth, especially at low temperature where the material does not flow, and "is thought to" reflect higher molecular mobility at the surface near or below Tg trasi2014a C4. The later ASD papers state the same as a widely reported failure mechanism, surface growth being faster than bulk growth by orders of magnitude duong2022c C2; moseson2023 C1. For felodipine, Kestur and Taylor also report a jump in growth below 50 °C for both surface and bulk crystals, "most likely" a glass–crystal (diffusionless) growth mode activated near Tg kestur2013a C3. Where such modes occur, extrapolating high-temperature growth rates to storage temperatures "may not be predictive" and, without the surface-enhanced mode, "an over prediction of the stability" may result kestur2013a C7, C18. Drug at the surface of an ASD is also more exposed to water and pH, for example in acid where an enteric polymer is a poor inhibitor ("can be attributed") nguyen2023b C37. During dissolution, crystallization is thought to start at the surface of amorphous solids, which a dissolved polymer may delay by interacting with that surface ("reasonable to speculate") alonzo2010 C36. (synthesis: the 2010 storage test and the 2014 bulk growth rates used sandwiched geometries, so their rankings refer to bulk behaviour; for powders, films and particles exposed to air or water the surface data above suggest faster crystallization, by about a factor of 6 for felodipine at 30 °C baird2010 C45; trasi2014a P1; kestur2013b R12.)

Conditions and limits

Surface and bulk growth were measured in different T/Tg windows in the 14-drug comparison, so most of those rank comparisons span different temperatures trasi2014a X16. The felodipine surface–bulk data are for one polymorph of one drug kestur2013a R8. The nilutamide surface advantage is stated without numbers trasi2014b R1. The coating studies concern polymer dispersions at high humidity, not neat drugs duong2022c R16; moseson2023 R27.

Counterexamples and conflicts

Above about 70 °C felodipine surface and bulk growth did not differ kestur2013a R9. Loratadine was slowest both at the surface and in the bulk trasi2014a C8. None found against faster surface crystallization near and below Tg.

Transfer check

Test open-surface films or powders as well as sandwiched films, at storage-relevant temperatures near or below Tg, where the surface advantage appears kestur2013a R9; kestur2013b R12. Inspect particle and tablet surfaces during stability and dissolution studies, for example by SEM, which revealed surface-grown crystals in these studies duong2022c R16; correasoto2017 C26. For high-load spray-dried dispersions that crystallize at the surface within weeks, a surface barrier such as an alumina coating is the strategy the library has tested duong2022c C33, C35.

History

The 2010 storage test assumed sandwiched films suppress surface crystallization baird2010 C45. Kestur in 2013 quantified surface-enhanced growth of felodipine and warned that bulk high-temperature data can overpredict stability kestur2013b R12; kestur2013a C18. Trasi in 2014 compared surface and bulk ranks for 14 drugs and relied on surface nucleation for nilutamide trasi2014a C8; trasi2014b R1. Surface coatings from 2022 targeted this failure mode duong2022c C2; moseson2022a R11; moseson2023 C1.

01Development line

One entry per paper in year order: what the library already knew, what the paper added, and how it relates to earlier entries.

  1. 2007
    towler2007solution interactions and crystallization outcomefirst proposed

    Spectroscopic Characterization of Intermolecular Interactions in Solution and Their Influence on Crystallization Outcome

    Known before
    IR and Raman band assignments of α, γ and amorphous indomethacin, taken from the 1997 PVP–indomethacin study (builds_on).
    Added
    concentration-dependent IR and Raman spectra of anthranilic acid in five solvents and indomethacin in two, compared with the polymorph that crystallized towler2007 R6, R13.
    Relation
    first proposed in this chapter that solution motifs are "not necessarily indicative" of the polymorph crystallized first towler2007 C33, C35.
    SupportsF8
  2. 2008
    towler2008counterion and amorphous organic saltsextends

    An Investigation into the Influence of Counterion on the Properties of Some Amorphous Organic Salts

    Known before
    counterion-dependent Tg of amorphous indomethacin salts (2002 Tong) and weight loss during sorption read as crystallization (2007 Gift) (builds_on).
    Added
    Tg, crystallization on heating and moisture behaviour of propranolol and nicardipine salts with many counterions, and a PLS model of Tg towler2008 R4, R5, R12, R17.
    Relation
    extends the salt Tg result to two bases; every salt raised Tg towler2008 C7, but the counterion effect on crystallization tendency needed "more work" towler2008 C34.
    SupportsF20
  3. 2010
    baird2010DSC classification from undercooled meltsfirst proposed

    A Classification System to Assess the Crystallization Tendency of Organic Molecules from Undercooled Melts

    Known before
    no earlier library paper; the methods came from inorganic glass science (Barandiaran–Colmenero and Cabral critical cooling rates, glass-stability parameters, reduced Tg, fragility) (builds_on).
    Added
    a heat–cool–heat DSC screen of 51 molecules into classes I, II and III, with a liquid-N2 quench splitting class I baird2010 R1, R3, C8, tests of rate robustness baird2010 C13, of critical-rate extrapolation baird2010 C20, of Trg, glass-stability and fragility parameters baird2010 C23, C31, a descriptor PCA baird2010 R20 and storage of glasses baird2010 R23, R24, R25.
    Relation
    first proposed; offered as a tool for early development baird2010 C64.
  4. 2010
    vaneerdenbrugh2010aclassification after rapid solvent evaporationconfirmsextends

    Crystallization Tendency of Active Pharmaceutical Ingredients Following Rapid Solvent Evaporation—Classification and Comparison with Crystallization Tendency from Undercooled Melts

    Known before
    the 51 compounds, their melt classes and DSC descriptors of baird2010 (builds_on).
    Added
    spin-coated films of 40 drugs from three solvents, sorted into three classes, with slow evaporation as a harsher test vaneerdenbrugh2010a R9, R13.
    Relation
    confirms the melt classes in 68% of drug–solvent combinations, with 30% one class apart vaneerdenbrugh2010a R16; extends the descriptor analysis to class models for a first-pass prediction vaneerdenbrugh2010a C46.
    SupportsF6F9F13F14
  5. 2012
    baird2012melt viscosity and glass-forming abilityconfirmsextends

    Role of Viscosity in Influencing the Glass-Forming Ability of Organic Molecules from the Undercooled Melt State

    Known before
    the classes, Tg values and compound set of baird2010 (builds_on).
    Added
    melt viscosity from Tm down to crystallization for 34 compounds, its rise on cooling, and VTF fits baird2012 R1, R9, R12.
    Relation
    extends baird2010 with a kinetic explanation: about 10⁻² Pa·s at Tm appeared as a threshold between glass formers and non-glass formers baird2012 C46, viscosity being "one critical attribute" baird2012 C48; it confirms baird2010 that kinetic fragility does not separate the classes baird2012 C43.
    SupportsF5F12
  6. 2013
    baird2013rapid heat–cool calorimetryextendsrevises

    Evaluating the non-isothermal crystallization behavior of organic molecules from the undercooled melt state using rapid heat/cool calorimetry

    Known before
    the classes, conventional-DSC data and the quenched-in nuclei proposal of baird2010 (builds_on).
    Added
    cooling and heating at up to about 2000 °C/min, critical cooling rates fitted over a wide rate range and critical heating rates of class II glasses baird2013 P2, R1, R7, R9.
    Relation
    revises the baird2010 verdict on critical-rate extrapolation in part, since a wider rate range "seems to improve" the fits baird2013 C20; extends the quenched-in nuclei proposal with the cooling–heating asymmetry, reported as a first for organic systems baird2013 C26, C30.
  7. 2014
    trasi2014acrystal growth rates of 31 drug liquidsextendscontradicts

    Factors Influencing Crystal Growth Rates from Undercooled Liquids of Pharmaceutical Compounds

    Known before
    the classes of baird2010, the melt viscosities of baird2012 and the separate polymer effects on nucleation and growth of acetaminophen (trasi2012a) (builds_on).
    Added
    seeded growth rates of 31 drugs, spanning 4 orders of magnitude in maximum velocity trasi2014a C23, their relation to viscosity, Tm − Tg and melting entropy trasi2014a R9, R11, PVP effects trasi2014a R6 and an extended nucleation-and-growth classification trasi2014a C17.
    Relation
    extends the DSC classes with growth; contradicts a reading of class III as slow-growing, since good glass formers include some of the fastest growers trasi2014a R1, C14.
  8. 2014
    trasi2014bnilutamide nucleation far below Tgconfirmscontradicts

    Nucleation and crystal growth of amorphous nilutamide – unusual low temperature behavior

    Known before
    nilutamide as a class III glass former (baird2010), flutamide and acetaminophen crystallizing on reheating (baird2013), and faster high-temperature growth of stable felodipine forms (kestur2013a) (builds_on).
    Added
    nucleation mainly far below Tg, a new metastable form I, polymorph growth-rate crossover and polymer effects on growth trasi2014b R5, R9, R16, R8, R3.
    Relation
    contradicts the general expectation that low-temperature storage stabilizes amorphous systems trasi2014b C57 and differs from flutamide and acetaminophen, which nucleate above Tg trasi2014b C37; confirms for one drug that good glass formation can come from low nucleation with fast growth trasi2014b C9.
    SupportsF3F4F7F15F19
  9. 2016
    lu2016vemurafenib polymorphs from the supercooled liquidconfirmsextends

    Vemurafenib: A Tetramorphic System Displaying Concomitant Crystallization from the Supercooled Liquid

    Known before
    the preliminary maximum crystal velocity of vemurafenib and the comparison set of trasi2014a (builds_on).
    Added
    four polymorphs in six morphologies, a temperature map of which forms appear, growth rates per form and their conversions lu2016 C41, C59, R12.
    Relation
    confirms the trasi2014a growth maximum near 0.94 Tm within 10 °C lu2016 C18 and that vemurafenib grows very rapidly lu2016 C19; extends the chapter to polymorph selection, where the stable form's larger driving force did not decide the outcome lu2016 C52.
    SupportsF1F2F3F7F9
For the authors: coverage of each paper card's main findings
paperfinding of section 0.3used in
towler2007F1. Every solvent gives a different anthranilic acid spectrum, yet form II always crystallizesF8
towler2007F2. In toluene the NH2 bands match form III, not form IIF8
towler2007F3. In acetonitrile a dimer band that matches form II grows with concentrationF8
towler2007F4. Indomethacin forms carboxylic acid dimers as concentration rises, in both solvents and even below saturationF8
towler2007F5. In ethanol the solution grows more γ-like, yet α crystallizesF8
towler2007F6. The acid–amide hydrogen bond of α is absent from the solutions that give αF8
towler2008F1. Every salt has a higher Tg than its free baseF20
towler2008F2. Tg falls roughly as counterion pKa rises, with exceptionsF20
towler2008F3. Amorphization and crystallization depend on counterion and on the kind of stressF20
towler2008F4. Moisture uptake varies widely with counterion, with no pKa linkF20
towler2008F5. A PLS model on counterion descriptors fits Tg within 3–4 °CF20
towler2008F6. pKa, chemical potential and electrophilicity weigh mostnot used: the loadings explain the Tg model, not crystallization; F20 uses the Tg data and the model fit only
baird2010F1. One DSC cycle splits the 51 molecules into three classesF9
baird2010F2. Liquid-N2 quenching splits class (I) in twoF9, F15
baird2010F3. Most class (II) and (III) assignments survive a change of scan rateF10
baird2010F4. Critical cooling rates fitted from DSC data were unreliableF11
baird2010F5. Trg, glass-stability parameters and fragility did not track the classesF12
baird2010F6. Molecular descriptors separate class (I) from class (III)F13, F6
baird2010F7. In stored glasses, crystallization speed followed the classF17, F15
vaneerdenbrugh2010aF1. Drugs fall into three classes, and the solvent rarely mattersF14, F9
vaneerdenbrugh2010aF2. Slow evaporation crystallizes most slow crystallizersF14
vaneerdenbrugh2010aF3. The solvent class matches the melt class in 68% of casesF14
vaneerdenbrugh2010aF4. PCA scores partly separate rapid from slow crystallizersF13
vaneerdenbrugh2010aF5. Slow crystallizers are larger, more flexible and higher in TgF13, F6
vaneerdenbrugh2010aF6. Class models give a first-pass prediction, checked on three drugsF13
baird2012F1. Fast crystallizers have runny melts, slow crystallizers viscous onesF5
baird2012F2. Viscosity of slow crystallizers rises faster on coolingF5
baird2012F3. Within a class, viscosity does not rank crystallizationF5
baird2012F4. Similar Tg/Tm, different viscosity rise and crystallizationF12
baird2012F5. Every fitted liquid is fragile, whatever its classF12
baird2013F1. The very-high-tendency compounds crystallize even at the fastest coolingF9
baird2013F2. The "high" class splits into compounds that need 25 °C/min and ones that crystallize even at 687–1220 °C/minF10
baird2013F3. Regression estimates put Rcrit,cool at a few thousand K/min where the method worksF11
baird2013F4. Class (II) glasses differ by four orders of magnitude in the reheating rate they needF10
baird2013F5. Reheating needs far faster rates than coolingF15
baird2013F6. Faster prior cooling lowers the reheating rate needed for many compoundsF16
trasi2014aF1. Growth rates differ by about four orders of magnitude between drugsF1
trasi2014aF2. Good glass formers include some of the fastest growersF15
trasi2014aF3. Slow growers have viscous melts and a small Tm − TgF5
trasi2014aF4. Viscosity alone does not set growth across compoundsF5
trasi2014aF5. Melting entropy explains part of the remaining spreadF6
trasi2014aF6. PVP slowed fast and slow growers by a similar factorF19
trasi2014bF1. Crystals grow fast, yet the melt forms a glassF15, F3, F13
trasi2014bF2. Nucleation happens mainly far below TgF4
trasi2014bF3. The low-temperature nuclei are a new metastable formF4, F7
trasi2014bF4. The two forms swap growth speed between 60 and 70 °CF7
trasi2014bF5. Form I has a different hydrogen-bond patternnot used: spectroscopic identification of form I, which bears on form identity rather than crystallization tendency
trasi2014bF6. The stable crystal holds a higher-energy conformer; form I and the glass differ from itF7 (conditions and limits: the conformer explanation is cited there as untested)
trasi2014bF7. PVP binds nilutamide and slows its growth; PAA does neitherF19
lu2016F1. The melt crystallizes readily on cooling; the glass crystallizes on heating in several stepsF9
lu2016F2. Four polymorphs in six morphologiesF7
lu2016F3. Which forms appear depends on the crystallization temperatureF7
lu2016F4. Growth rates differ by up to two orders of magnitudeF2, F1
lu2016F5. The metastable forms convert step by step: γ and δ to β, β to αF7
lu2016F6. α is the stable form and is monotropically related to βF7

01Concepts

crystallization tendencythe property the chapter is aboutcrystallization tendency classificationDSC and spin-coating classes I, II, III used by every later paperextended nucleation and growth classificationclasses IA–IV combining DSC behaviour with growth rateglass-forming abilityease of vitrifying the melt on coolingglass stabilityresistance of the glass to crystallization on reheating or storageglass-stability parametersinorganic-glass indices tested and rejected for organicscrystallization from the supercooled meltthe experimental route of most paperssupercooled liquidthe state between Tm and Tg in which nucleation and growth are measuredcold crystallizationcrystallization of the glass on reheating, the class II signaturenucleationthe step that separates class III good glass formers from fast crystallizersnucleation temperature zonetemperature window of nucleation relative to growthsub-Tg nucleationnucleation far below Tg (nilutamide)heterogeneous nucleationpan, plate and surface nucleation that limit the screensquenched-in nucleiproposed cause of the cooling-rate effect on reheatingclassical nucleation theoryframework for the driving-force and viscosity argumentsprenucleation clusterssolute association before nucleation, tested against polymorph outcomecrystal growth ratemeasured growth from the melt, 4 orders between drugsmaximum crystal growth velocitythe per-drug maximum used to group growerscrystal growthgrowth as a process, including surface and polymer effectscrystal growth inhibition by polymersreduction of melt growth by PVP and other polymerssurface crystallizationfaster growth at free surfacesmelt viscositymobility term that separates the extreme classesfragilitycurvature of viscosity on cooling; did not separate classesglass transition temperaturereference temperature for storage, Trg and Tm − Tgreduced glass transition temperatureTg/Tm, tested as a glass-forming criterionTm − Tg temperature gaprange to cross on cooling; tracked growth groupsdegree of supercoolingundercooling at crystallization and the driving-force variablethermodynamic driving force for crystallizationfree energy difference between liquid and crystalHoffman equationestimate of the driving force from melting dataenthalpy of fusiondescriptor and Hoffman inputentropy of fusiondescriptor and growth-rate factorconformational flexibilitymolecular descriptor of slow crystallizerscritical cooling rateslowest cooling that avoids crystallizationcritical heating rateslowest reheating of a glass that avoids crystallizationBarandiaran-Colmenero methodextrapolation of the critical cooling rate from DSCsolvent evaporation ratevariable of the spin-coating against slow-evaporation comparisonpolymorphismseveral forms from one supercooled liquidmonotropystability relation of the polymorph pairs studiedBurger-Ramberger rulesbasis of the monotropy assignmentsphysical stabilitythe formulation outcome the classes are meant to flagamorphous saltcounterion effects on Tg and crystallization

02Formulas

Hoffman equation\Delta G_v = -\frac{\Delta H_{fus}\,(T_{melt,ons} - T)\,T}{T_{melt,ons}^{2}}driving force as a descriptor and for polymorph pairsclassical nucleation barrierW^* = \frac{16\pi\gamma^3}{3\,\Delta G_v^2}argument linking driving force to nucleationtemperature of maximum crystal growth rateT_{cryst,max} \approx 0.94\,T_m0.94 Tm rule checked against measured maximaentropy of fusion\Delta S_{fus} = \frac{\Delta H_{fus}}{T_{melt,ons}}descriptor in both class PCAsBarandiaran-Colmenero critical cooling rate\ln(\text{rate}) = A - \frac{B}{(\Delta T)^2},\quad \Delta T = T_m - T_{crys,max}critical cooling rate from undercooling at several ratesCabral critical cooling rateR_{crit} = \exp\left(A^* - \frac{B^*}{T_m^2}\right)corrected critical cooling rateBurger-Ramberger heat-of-fusion ruleNot printed in either paper: applied in words from the relative melting points and enthalpies of fusionmonotropy of nilutamide and vemurafenib formsVogel-Tammann-Fulcher equation\tau = \tau_0 \exp\left(\frac{D\,T_0}{T - T_0}\right)strength parameter and fragilityreduced glass transition temperatureT_{rg} = \frac{T_g}{T_{melt}}Tg/Tm criterion tested against classescritical cooling rate to avoid nucleationR_{Ncrit} \sim \frac{1}{\tau_{Nmin}}argument for the nucleation windowlinear fit of melt viscosity on Tm − Tgy = -0.0028\,x + 0.3989,\quad R^2 = 0.7368trend between viscosity and Tm − Tglinear fit of normalized growth rate on melting entropyy = -0.6\,x - 0.95,\quad R^2 = 0.61entropy effect at equal mobilityJackson interface-kinetics growth rateU = \frac{2 k_B T}{\pi \lambda^2 \eta}\, f \left[1 - \exp\left(-\frac{\Delta H\,\Delta T}{k_B T\,T_m}\right)\right] \exp\left(-\frac{\Delta S}{k_B}\right)growth model behind the entropy analysis
39 more formulas printed by one paper (table only)
formularole hereexpression
degree of undercoolingundercooling at crystallization on cooling\Delta T = T_m - T_{cryst} baird2013 E4
Lu–Liu parameter KLLglass-stability parameterK_{LL} = \frac{T_{cryst,ons}}{T_{g,mid} + T_{melt}} baird2010 E5
Hruby parameter KHglass-stability parameterK_H = \frac{T_{cryst,ons} - T_{g,mid}}{T_{melt} - T_{cryst,ons}} baird2010 E6
Weinberg parameter KWglass-stability parameterK_W = \frac{T_{cryst,max} - T_{g,mid}}{T_{melt}} baird2010 E7
Saad–Poulain parameter KSPglass-stability parameterK_{SP} = \frac{(T_{cryst,ons} - T_{g,mid})(T_{cryst,max} - T_{cryst,ons})}{T_{g,mid}} baird2010 E8
Nascimento parameter K1glass-stability parameterK_1 = \frac{T_{cryst,ons}}{T_{melt}} baird2010 E9
Nascimento parameter K2glass-stability parameterK_2 = \frac{(T_{cryst,ons} - T_{g,mid})(T_{cryst,max} - T_{cryst,ons})}{T_{melt}} baird2010 E10
reduced temperature Tredglass-stability parameterT_{red} = \frac{T_{cryst,ons} - T_{g,mid}}{T_{melt} - T_{g,mid}} baird2010 E11
fragility indexdefinition of mm = \frac{d \log \tau}{d(T_g/T)} at T = T_g baird2010 E13
fragility from activation enthalpym of class II and IIIm = \frac{\Delta E_{T_g}}{(\ln 10)\,R\,T_g} baird2010 E14
Moynihan heating-rate dependence of Tgactivation enthalpy from Tg shifts\frac{d(\ln q)}{d(1/T_g)} = -\frac{\Delta E_{T_g}}{R} baird2010 E15
fragility–strength relationlink of m and Dm = \frac{D\,T_0/T}{(\ln 10)\,(1 - T_0/T_g)^2} baird2010 E16
minimum fragilitym_{min} = 16m_{min} = \log\left(\tau(T_g)/\tau_0\right) baird2010 E17
strength parameter from fragilityD^aD = \frac{\ln(10)\,m_{min}^2}{m - m_{min}} baird2010 E18
Kauzmann temperature from configurational heat capacityTK for D^b\frac{1}{T_K} = \frac{1}{T_{melt}}\left(1 + \frac{\Delta H_{fus}}{C_{p,conf}\,T_g}\right) baird2010 E19
configurational to calorimetric heat capacity ratioinput of E19\frac{\Delta C_p}{C_{p,conf}} \approx 0.9 baird2010 E20
nucleation time and nucleation rateargument only\tau_N = \frac{1}{J_N V} baird2010 E22
enthalpy and entropy difference below the melting pointdriving-force argument\Delta H(T) = \Delta H_{fus} - \int_T^{T_{melt}} \Delta C_p\,\delta T; \Delta S(T) = \Delta S_{fus} - \int_T^{T_{melt}} \frac{\Delta C_p}{T}\,\delta T baird2010 E23
linear regression of reduced Tg on molecular weightTg estimate for 16 class I moleculesT_{rg} = 0.00021\,\mathrm{MW} + 0.64433, R² 0.44204 baird2010 E27
prevalence of a class pairspin-coating against melt class\text{percentage} = 100 \times \frac{\text{combinations in the group}}{93} vaneerdenbrugh2010a E4
weight-to-volume basis conversiondescriptors per volume\text{value per volume} = \text{weight-based value} \times \text{crystalline density} vaneerdenbrugh2010a E5
film thickness from weight gainspin-coated film thicknessthickness from substrate weight gain with 1.28 g/mL vaneerdenbrugh2010a E6
Tg from the Tg/Tm ratioTg of fast crystallizersTg predicted from Tg/Tm ratios of 35 compounds vaneerdenbrugh2010a E7
steady-state homogeneous nucleation rateviscosity argumentI_{SS} = I_0 \exp\left[-\frac{W^* + \Delta G_\eta}{k_B T}\right] baird2012 E1
screw-dislocation growth rateviscosity argumentU = f\,\frac{k_B T}{\lambda^2 \eta}\left[1 - \exp\left(\frac{\Delta G_v}{k_B T}\right)\right] baird2012 E3
VTF relation between T0, D and Tgcheck of fitted T0\frac{T_g}{T_0} = 1 + \frac{D}{2.303\,\log(\eta_g/\eta_0)} baird2012 E5
viscosity ratiorise of viscosity on undercooling\eta/\eta_{melt} baird2012 E7
kinetic growth coefficientgrowth corrected for driving forceU_{kin} = \frac{U}{1 - e^{-\Delta G/RT}} trasi2014a E2
driving force from melting entropyΔG estimate\Delta G \approx \Delta S_m\,\Delta T trasi2014a E3
structural relaxation time from viscositynormalization of growth\tau_\alpha = \frac{C\eta}{G} trasi2014a E4
standardized deviation of a group averageproperty profile of growth groups\frac{\text{Average}_{Group} - \text{Average}_{All}}{\text{Stdev}_{All}} trasi2014a E5
linear growth rate from length–time slopenilutamide growth rates\text{growth rate} = \text{slope of crystal length against time} trasi2014b E1
free energy difference between polymorphsnilutamide form I against stable formrelation not printed trasi2014b E3
supersaturation ratioindomethacin solutions\text{supersaturation} = c/c^* towler2007 E1
chemical hardnesscounterion descriptor\eta \cong I - A towler2008 E1
electronic chemical potentialcounterion descriptor\mu = -\frac{I + A}{2} towler2008 E2
electrophilicity indexcounterion descriptor\omega = \mu^2/2\eta towler2008 E3
chemical softnesscounterion descriptorS = \frac{1}{\eta} towler2008 E4
partial least squares regressionTg model of the saltsPLS2 (NIPALS), not printed towler2008 E5

Sources: Crystallization tendency

Every paper cited in the chapter Crystallization tendency (category Solid state stability) and in the concept and formula pages drafted from it. Keys are first-author surname plus year; a, b, c separate papers of the same author and year. The first nine rows are the papers of the subcategory.

keyyeartitleslugcategory / subcategory
towler20072007Spectroscopic Characterization of Intermolecular Interactions in Solution and Their Influence on Crystallization Outcome2007-towler-solution-interactions-crystallization-outcomeSolid state stability / Crystallization tendency
towler20082008An Investigation into the Influence of Counterion on the Properties of Some Amorphous Organic Salts2008-towler-counterion-amorphous-organic-saltsSolid state stability / Crystallization tendency
baird20102010A Classification System to Assess the Crystallization Tendency of Organic Molecules from Undercooled Melts2010-baird-classification-crystallization-tendency-undercooled-meltsSolid state stability / Crystallization tendency
vaneerdenbrugh2010a2010Crystallization Tendency of Active Pharmaceutical Ingredients Following Rapid Solvent Evaporation—Classification and Comparison with Crystallization Tendency from Undercooled Melts2010-van-eerdenbrugh-api-crystallization-tendency-rapid-solvent-evaporationSolid state stability / Crystallization tendency
baird20122012Role of Viscosity in Influencing the Glass-Forming Ability of Organic Molecules from the Undercooled Melt State2012-baird-viscosity-glass-forming-ability-undercooled-meltsSolid state stability / Crystallization tendency
baird20132013Evaluating the non-isothermal crystallization behavior of organic molecules from the undercooled melt state using rapid heat/cool calorimetry2013-baird-rapid-heat-cool-crystallization-undercooled-melt-glass-formingSolid state stability / Crystallization tendency
trasi2014a2014Factors Influencing Crystal Growth Rates from Undercooled Liquids of Pharmaceutical Compounds2014-trasi-crystal-growth-rates-undercooled-pharmaceutical-liquidsSolid state stability / Crystallization tendency
trasi2014b2014Nucleation and crystal growth of amorphous nilutamide – unusual low temperature behavior2014-trasi-taylor-amorphous-nilutamide-low-temperature-nucleationSolid state stability / Crystallization tendency
lu20162016Vemurafenib: A Tetramorphic System Displaying Concomitant Crystallization from the Supercooled Liquid2016-lu-vemurafenib-tetramorphic-cold-crystallization-supercooled-liquidSolid state stability / Crystallization tendency
taylor19971997Spectroscopic Characterization of Interactions Between PVP and Indomethacin in Amorphous Molecular Dispersions1997-taylor-pvp-indomethacin-amorphous-molecular-dispersionsSolid state stability / Miscibility and drug-polymer interactions
marsac2006a2006A Comparison of the Physical Stability of Amorphous Felodipine and Nifedipine Systems2006-marsac-physical-stability-amorphous-felodipine-nifedipineSolid state stability / Physical stability
marsac2006b2006Theoretical and Practical Approaches for Prediction of Drug–Polymer Miscibility and Solubility2006-marsac-drug-polymer-miscibility-solubilitySolid state stability / Miscibility and drug-polymer interactions
konno20062006Influence of Different Polymers on the Crystallization Tendency of Molecularly Dispersed Amorphous Felodipine2006-konno-felodipine-polymer-crystallization-nucleation-solid-dispersionsSolid state stability / Physical stability
konno20082008Ability of Different Polymers to Inhibit the Crystallization of Amorphous Felodipine in the Presence of Moisture2008-konno-polymers-inhibit-amorphous-felodipine-moisture-crystallizationSolid state stability / Physical stability
marsac2008a2008Recrystallization of Nifedipine and Felodipine from Amorphous Molecular Level Solid Dispersions Containing Poly(vinylpyrrolidone) and Sorbed Water2008-marsac-nifedipine-felodipine-pvp-sorbed-water-recrystallizationSolid state stability / Physical stability
marsac2008b2008Spontaneous Crystallinity Loss of Drugs in the Disordered Regions of Poly(Ethylene Oxide) in the Presence of Water2008-marsac-peo-water-spontaneous-drug-crystallinity-lossSolid state stability / PEG dispersions
marsac20092009Estimation of Drug–Polymer Miscibility and Solubility in Amorphous Solid Dispersions Using Experimentally Determined Interaction Parameters2009-marsac-drug-polymer-miscibility-solubility-asd-interaction-parametersSolid state stability / Miscibility and drug-polymer interactions
alonzo20102010Understanding the Behavior of Amorphous Pharmaceutical Systems during Dissolution2010-alonzo-amorphous-pharmaceutical-dissolution-crystallization-polymersDissolution / B. Supersaturation from ASD dissolution
kestur2010a2010Role of polymer chemistry in influencing crystal growth rates from amorphous felodipine2010-kestur-polymer-chemistry-felodipine-crystal-growth-inhibitionSolid state stability / Miscibility and drug-polymer interactions
kestur2010b2010Effects of the Molecular Weight and Concentration of Polymer Additives, and Temperature on the Melt Crystallization Kinetics of a Small Drug Molecule2010-kestur-pvp-mw-concentration-temperature-felodipine-melt-crystallizationSolid state stability / Physical stability
marsac20102010Effect of Temperature and Moisture on the Miscibility of Amorphous Dispersions of Felodipine and Poly(vinyl pyrrolidone)2010-marsac-temperature-moisture-miscibility-felodipine-pvp-asdSolid state stability / Physical stability
rumondor20102010Effects of Moisture on the Growth Rate of Felodipine Crystals in the Presence and Absence of Polymers2010-rumondor-moisture-felodipine-crystal-growth-pvp-hpmcasSolid state stability / Physical stability
vaneerdenbrugh2010b2010Small Scale Screening To Determine the Ability of Different Polymers To Inhibit Drug Crystallization upon Rapid Solvent Evaporation2010-van-eerdenbrugh-polymer-crystallization-inhibition-spin-coating-screenSolid state stability / Miscibility and drug-polymer interactions
kestur20112011Influence of polymer chemistry on crystal growth inhibition of two chemically diverse organic molecules2011-kestur-polymer-chemistry-crystal-growth-bifonazole-nimesulideSolid state stability / Miscibility and drug-polymer interactions
zhu20112011Time-Resolved SAXS/WAXS Study of the Phase Behavior and Microstructural Evolution of Drug/PEG Solid Dispersions2011-zhu-time-resolved-saxs-waxs-drug-peg-solid-dispersionsSolid state stability / PEG dispersions
hsieh20122012pH-Induced Precipitation Behavior of Weakly Basic Compounds: Determination of Extent and Duration of Supersaturation Using Potentiometric Titration and Correlation to Solid State Properties2012-hsieh-ph-induced-precipitation-weak-bases-supersaturation-solid-stateDissolution / D. Crystallization from solution
trasi2012a2012Effect of polymers on nucleation and crystal growth of amorphous acetaminophen2012-trasi-taylor-polymers-nucleation-growth-amorphous-acetaminophenSolid state stability / Physical stability
trasi2012b2012Effect of Additives on Crystal Growth and Nucleation of Amorphous Flutamide2012-trasi-flutamide-additives-growth-nucleationSolid state stability / Physical stability
zhu20122012Modification of Crystallization Behavior in Drug/Polyethylene Glycol Solid Dispersions2012-zhu-api-peg-solid-dispersion-crystallization-behaviorSolid state stability / PEG dispersions
ilevbare2013a2013Liquid-Liquid Phase Separation in Highly Supersaturated Aqueous Solutions of Poorly-Water Soluble Drugs – Implications for Solubility Enhancing Formulations2013-ilevbare-llps-supersaturated-poorly-water-soluble-drugsDissolution / C. Amorphous solubility and LLPS
ilevbare2013b2013Impact of Polymers on Crystal Growth Rate of Structurally Diverse Compounds from Aqueous Solution2013-ilevbare-polymers-crystal-growth-structurally-diverse-compoundsDissolution / D. Crystallization from solution
ilevbare2013c2013Maintaining Supersaturation in Aqueous Drug Solutions: Impact of Different Polymers on Induction Times2013-ilevbare-polymers-supersaturation-induction-timesDissolution / D. Crystallization from solution
kestur2013a2013Evaluation of the crystal growth rate of felodipine polymorphs in the presence and absence of additives as a function of temperature2013-kestur-felodipine-polymorph-crystal-growth-pvp-additivesSolid state stability / Physical stability
kestur2013b2013Influence of particle size on the crystallization kinetics of amorphous felodipine powders2013-kestur-particle-size-crystallization-amorphous-felodipine-pvpSolid state stability / Physical stability
wegiel2013a2013Mid-infrared spectroscopy as a polymer selection tool for formulating amorphous solid dispersions2013-wegiel-mid-ir-polymer-selection-polyphenol-asd-stabilitySolid state stability / Miscibility and drug-polymer interactions
wegiel2013b2013Crystallization of Amorphous Solid Dispersions of Resveratrol during Preparation and Storage—Impact of Different Polymers2013-wegiel-resveratrol-asd-polymer-crystallization-stabilitySolid state stability / Physical stability
hsieh20142014Assessing the Impact of Polymers on the pH-Induced Precipitation Behavior of Poorly Water-Soluble Compounds using Synchrotron Wide Angle X-Ray Scattering2014-hsieh-polymers-ph-induced-precipitation-waxsDissolution / D. Crystallization from solution
mosqueragiraldo20142014Impact of surfactants on the crystal growth of amorphous celecoxib2014-mosquera-giraldo-surfactants-crystal-growth-amorphous-celecoxibSolid state stability / Physical stability
raina2014a2014Enhancements and Limits in Drug Membrane Transport Using Supersaturated Solutions of Poorly Water Soluble Drugs2014-raina-llps-limits-drug-membrane-transport-supersaturated-solutionsBioperformance / Bioperformance
raina2014b2014Impact of Polymers on the Crystallization and Phase Transition Kinetics of Amorphous Nifedipine during Dissolution in Aqueous Media2014-raina-polymers-nifedipine-crystallization-phase-transition-dissolutionDissolution / B. Supersaturation from ASD dissolution
wegiel20142014Curcumin amorphous solid dispersions: the influence of intra and intermolecular bonding on physical stability2014-wegiel-curcumin-asd-intramolecular-bonding-physical-stabilitySolid state stability / Physical stability
almeidaesousa20152015Assessment of the Amorphous “Solubility” of a Group of Diverse Drugs Using New Experimental and Theoretical Approaches2015-almeida-sousa-amorphous-solubility-diverse-drugs-llps-cp-methodDissolution / C. Amorphous solubility and LLPS
chen20152015Bile Salts as Crystallization Inhibitors of Supersaturated Solutions of Poorly Water-Soluble Compounds2015-chen-bile-salts-crystallization-inhibitors-supersaturated-solutionsDissolution / D. Crystallization from solution
indulkar20152015pH-Dependent Liquid−Liquid Phase Separation of Highly Supersaturated Solutions of Weakly Basic Drugs2015-indulkar-ph-dependent-llps-weak-bases-amorphous-solubilityDissolution / C. Amorphous solubility and LLPS
mosqueragiraldo20152015Glass–Liquid Phase Separation in Highly Supersaturated Aqueous Solutions of Telaprevir2015-mosquera-giraldo-telaprevir-glass-liquid-phase-separationDissolution / C. Amorphous solubility and LLPS
raina2015a2015Using Environment-Sensitive Fluorescent Probes to Characterize Liquid-Liquid Phase Separation in Supersaturated Solutions of Poorly Water Soluble Compounds2015-raina-fluorescent-probes-llps-supersaturated-solutionsCharacterization methods / Solution methods
raina2015b2015Trends in the Precipitation and Crystallization Behavior of Supersaturated Aqueous Solutions of Poorly Water-Soluble Drugs Assessed Using Synchrotron Radiation2015-raina-precipitation-crystallization-supersaturated-dihydropyridines-synchrotronDissolution / D. Crystallization from solution
trasi20152015Evaluating the influence of polymers on nucleation and growth in supersaturated solutions of acetaminophen2015-trasi-polymers-nucleation-growth-supersaturated-acetaminophenDissolution / D. Crystallization from solution
wegiel20152015Phase Behavior of Resveratrol Solid Dispersions Upon Addition to Aqueous media2015-wegiel-resveratrol-solid-dispersions-aqueous-phase-behaviorDissolution / B. Supersaturation from ASD dissolution
xie20152015Dissolution Performance of High Drug Loading Celecoxib Amorphous Solid Dispersions Formulated with Polymer Combinations2015-xie-taylor-celecoxib-high-drug-loading-asd-polymer-combinationsDissolution / B. Supersaturation from ASD dissolution
sato20162016Acceleration of the crystal growth rate of low molecular weight organic compounds in supercooled liquids in the presence of polyhydroxybutylate2016-sato-phb-crystal-growth-supercooled-organic-compoundsSolid state stability / Physical stability
tian20162016Investigating the Correlation between Miscibility and Physical Stability of Amorphous Solid Dispersions Using Fluorescence-based Techniques2016-tian-indomethacin-polymer-miscibility-physical-stability-fluorescenceSolid state stability / Physical stability
trasi20162016Non-Sink Dissolution Behavior and Solubility Limit of Commercial Tacrolimus Amorphous Formulations2016-trasi-commercial-tacrolimus-asd-nonsink-amorphous-solubilityDissolution / B. Supersaturation from ASD dissolution
correasoto20172017Second harmonic generation microscopy as a tool for the early detection of crystallization in spray dried dispersions2017-correa-soto-shg-early-crystallization-spray-dried-asdsCharacterization methods / Solid-state methods
lu20172017Maintaining Supersaturation of Active Pharmaceutical Ingredient Solutions with Biologically Relevant Bile Salts2017-lu-bile-salts-telaprevir-supersaturationDissolution / D. Crystallization from solution
tian20172017Impact of Polymers on the Melt Crystal Growth Rate of Indomethacin Polymorphs2017-tian-polymers-melt-crystal-growth-indomethacin-polymorphsSolid state stability / Physical stability
trasi20172017Evaluation of the Crystallization Tendency of Commercially Available Amorphous Tacrolimus Formulations Exposed to Different Stress Conditions2017-trasi-commercial-amorphous-tacrolimus-crystallization-stressSolid state stability / Physical stability
moseson20182018The application of temperature-composition phase diagrams for hot melt extrusion processing of amorphous solid dispersions to prevent residual crystallinity2018-moseson-temperature-composition-phase-diagrams-hme-asd-residual-crystallinityASD process / ASD process
trasi20192019Amorphous solid dispersion formation via solvent granulation – A case study with ritonavir and lopinavir2019-trasi-solvent-granulation-ritonavir-lopinavir-asdASD process / ASD process
moseson20202020Application and Limitations of Thermogravimetric Analysis to Delineate the Hot Melt Extrusion Chemical Stability Processing Window2020-moseson-tga-hme-chemical-stability-processing-windowASD process / ASD process
saboo20202020Congruent release of drug and polymer from amorphous solid dispersions: insights into the role of drug-polymer hydrogen bonding, surface crystallization and glass transition2020-saboo-congruent-release-hbond-crystallization-tg-pvpva-asdDissolution / A. Release mechanism and LoC
trasi20202020Physical stability and release properties of lumefantrine amorphous solid dispersion granules prepared by a simple solvent evaporation approach2020-trasi-lumefantrine-asd-granules-solvent-evaporationSolid state stability / Physical stability
wilson20212021Interaction of Polymers with Enzalutamide Nanodroplets—Impact on Droplet Properties and Induction Times2021-wilson-enzalutamide-nanodroplets-polymer-interactions-induction-timesDissolution / D. Crystallization from solution
duong2022a2022Combining enabling formulation strategies to generate supersaturated solutions of delamanid: In situ salt formation during amorphous solid dispersion fabrication for more robust release profiles2022-duong-delamanid-in-situ-salt-asd-releaseDissolution / B. Supersaturation from ASD dissolution
duong2022b2022Phase Behavior and Crystallization Kinetics of a Poorly Water-Soluble Weakly Basic Drug as a Function of Supersaturation and Media Composition2022-duong-posaconazole-phase-behavior-crystallization-kinetics-mediaDissolution / D. Crystallization from solution
duong2022c2022Surface nanocoating of high drug-loading spray-dried amorphous solid dispersions by atomic layer coating: Excellent physical stability under accelerated storage conditions for two years2022-van-duong-surface-nanocoating-high-drug-loading-asd-alc-stabilityASD process / ASD process
hiew2022a2022Balancing Solid-State Stability and Dissolution Performance of Lumefantrine Amorphous Solid Dispersions: The Role of Polymer Choice and Drug−Polymer Interactions2022-hiew-lumefantrine-asd-stability-dissolution-polymer-choiceSolid state stability / Miscibility and drug-polymer interactions
hiew2022b2022Combining drug salt formation with amorphous solid dispersions – a double edged sword2022-hiew-combining-drug-salt-formation-asd-double-edged-swordDissolution / A. Release mechanism and LoC
indulkar20222022Role of Surfactants on Release Performance of Amorphous Solid Dispersions of Ritonavir and Copovidone2022-indulkar-surfactants-ritonavir-copovidone-asd-release-performanceDissolution / A. Release mechanism and LoC
moseson2022a2022Atomic Layer Coating to Inhibit Surface Crystallization of Amorphous Pharmaceutical Powders2022-moseson-atomic-layer-coating-surface-crystallization-asd-powdersASD process / ASD process
moseson2022b2022Formulation and processing strategies which underpin susceptibility to matrix crystallization in amorphous solid dispersions2022-moseson-processing-strategies-matrix-crystallization-bcl-pvpva-asdASD process / ASD process
moseson20232023Impact of Aluminum Oxide Nanocoating on Drug Release from Amorphous Solid Dispersion Particles2023-moseson-aluminum-oxide-nanocoating-drug-release-asd-particlesASD process / ASD process
nguyen2023a2023Fed- and Fasted-State Performance of Pretomanid Amorphous Solid Dispersions Formulated with an Enteric Polymer2023-nguyen-pretomanid-hpmcas-asd-fed-fasted-performanceBioperformance / Bioperformance
nguyen2023b2023Enteric coating of tablets containing an amorphous solid dispersion of an enteric polymer and a weakly basic drug: A strategy to enhance in vitro release2023-nguyen-enteric-coating-delamanid-hpmcp-asd-releaseDissolution / B. Supersaturation from ASD dissolution
yang20232023Drug Release from Surfactant-Containing Amorphous Solid Dispersions: Mechanism and Role of Surfactant in Release Enhancement2023-yang-surfactant-containing-asd-release-enhancementDissolution / A. Release mechanism and LoC
For the authors: card issues found while writing

Card issues: Crystallization tendency

Material problems found in cards while writing this chapter. The nine subcategory cards were also scanned for cited ids that no row of the same card defines; none were found.

paperidproblem
2017-xie-temperature-moisture-celecoxib-binary-ternary-asd-stability (outside this subcategory)year fieldThe card gives year: 2016 (DOI 10.1016/j.xphs.2016.06.017), while its slug begins with 2017, so the year field and the slug disagree. Because the year could not be settled from the card, this chapter does not cite the paper.
← Concepts and formulas

concept · as used in Crystallization tendency

accelerated stability testing

Storage at raised humidity and/or temperature to provoke and rank physical instability in less time than at the intended storage condition. In the library the concern is that high RH can change the mechanism, not only the rate, of crystallization of susceptible dispersions [rumondor2009c C52].

Findings

F16, F25.

Related concepts

related to moisture-induced phase separation, moisture-induced crystallization and physical stability.

Formulas

Arrhenius equation (temperature extrapolation; used here for nucleation, not for shelf life).

How it is measured or obtained

Storage over saturated salts at fixed RH (here 54–94% RH at room temperature) and read-out by IR, DSC, PXRD or microscopy [rumondor2009c P4; rumondor2009b P2, P3; marsac2010 P7].

Values in the library
materialvalue or classconditionspaper ids
pimozide–PVPpartial phase separation after 3 days54% RH[rumondor2009c R10, C4]
felodipine–PVP 30.3 wt%distinct separation after 1 week75% RHmarsac2010 R7, R8
felodipine with 25–50% PVP vs HPMCASindistinguishable at 0–52% RH in ≈490 days; 51–82% vs 9–28% crystallized at 75–93% RH22–25 °C[rumondor2009b R19, R9]
PEO compacts (outside)slow water uptake over months; partially crystalline polymers "may" defeat Arrhenius extrapolation94% RHmarsac2008b C40
curcumin dispersions25 °C/0% RH and 90 °C/0% or 60 °C/98% RH separate no polymersix conditionswegiel2014 R13, C23
felodipine–3% PVP powder≈8-fold faster start and ≈85% against ≈45% plateau at 50 against 30 °Cdry storagekestur2013b R17, C38

Evidence table: accelerated stability testing.

Pitfalls and disagreements

In susceptible systems "the route of crystallization may be altered at high RH" and need not represent lower RH [rumondor2009c C52]; the separation threshold depends on time and composition marsac2010 X9. Dry and moderate-RH data could not rank PVP against HPMCAS for felodipine within the method error [rumondor2009b X4]. The 2013 papers state that their results "highlight the difficulty" of choosing accelerated conditions wegiel2013b C41; kestur2013b C40.

← Concepts and formulas

concept · as used in Crystallization tendency

acid-base interaction

The transfer of a proton, or the formation of an ion pair, between an acidic and a basic group of drug and polymer, instead of a neutral hydrogen bond. In the papers of this chapter: the basic dimethylamino group of E100 with the weakly acidic OH groups of naringenin and quercetin (also resveratrol and curcumin); the carboxylic acids of HPMCAS, HPMCP, CAP and Eudragit L 100 with the tertiary amine of lumefantrine; and the strongly acidic sulfonic acid of PSSA with the amine or ring nitrogen of anthranilic acid, lidocaine and dibucaine [wegiel2013a C16, C17, C33; hiew2022a C25, C48; vaneerdenbrugh2011a C22, C38, C44]. The authors take ionic interactions to be "typically stronger" than hydrogen bonds, an assumption, and ascribe the unusual stability of E100 dispersions to them ("appear to be the reason") wegiel2013a C16, C35. For the enteric polymers the same contact is held to reduce hydration of the polymer carboxylates and so to slow release ("are postulated"; "is thought to") hiew2022a C53.

Findings

Miscibility and drug-polymer interactions: F21, F23, F24, F25, F26. Physical stability: F24, F34.

Related concepts

depends on pKa; contrasts with hydrogen bonding, halogen bonding and hydrogen-bond donor and acceptor groups; part of drug-polymer interactions; acts on crystallization inhibition and physical stability; an alternative where intramolecular hydrogen bonding limits hydrogen bonds wegiel2014 C44.

Formulas

free carboxylate fraction after ion pairing, Fox equation.

How it is measured or obtained

By the loss of the E100 amine bands at 2820 and 2770 cm⁻¹ with rising drug content wegiel2013a R6; by a protonated component of the N 1s XPS peak and by carboxylate bands at 1594–1560 and 1410–1343 cm⁻¹ in subtraction IR spectra hiew2022a R19, R20; by ATR-FTIR of melts at equimolar functional groups (spectra not shown) [vaneerdenbrugh2011a R8, R15, R17]; by colour change (curcumin with E100) wegiel2014 R10, C20: dark red for curcumin–E100 against orange for the others, read as a change of the drug's ionization state; by a dispersion Tg above both components wegiel2014 C41. For the weak base lumefantrine with acidic enteric polymers in the physical-stability papers, XPS of the N 1s signal gives the share of protonated drug, and ATR-FTIR shows the polymer carbonyl band moving up and a band near 1560 cm⁻¹ read as carboxylate ("likely") [trasi2020 P15, R9, C11, C12; bhujbal2021 P6, R6]. The pKa of each group is calculated or titrated to judge whether a proton can move [vaneerdenbrugh2011a P7; hiew2022a P17].

Values in the library
materialvalue or classconditionspaper ids
quercetin–E100amine bands gone at 50% quercetinIR, filmswegiel2013a R6
resveratrol–E100; curcumin–E100bands absent at 50% resveratrol; eradicated at 60% curcuminIR, filmswegiel2013b R7, C15; wegiel2014 R7, C16
lumefantrine, 25% drug loadingprotonated 19% (HPMCAS), 46% (HPMCP), 55% (CAP), 49% (Eudragit L 100); 0% with PVPVAXPS, dry compactshiew2022a R20
same, after 2 h in pH 6.8 buffer3, 21, 54, 12%XPShiew2022a R20
anthranilic acid, lidocaine, dibucaine with PSSAproton transfer reported; not with PAAATR-FTIR of melts (not shown)[vaneerdenbrugh2011a R8, R15, R17]
cationic drugs with HPMCAS-MF, 1:1 drug to carboxyl, pH 6.0≥70% of the polymer complexedco-precipitation[bapat2024 C19]
curcumin–E100 (basic monomer pKa 8.4; curcumin pKa 10.51, 9.88, 8.38)dark red; Tg 62 °C, about 19 °C above the Fox line after recalculation75:25 powderwegiel2014 M5, M1, R10, X1, C41
resveratrol–E100highest amorphous loading of six polymers (60%); "could be responsible" for better manufacturabilityrotary evaporationwegiel2013b R2, C16, C43
curcumin–E100; resveratrol–E100amorphous > 350 and > 400 days at 84% RHstorage, 25 °Cwegiel2014 R13; wegiel2013b R12.g
lumefantrine 40:60 with HPMCAS, Eudragit L100, HPMCP, CAP16, 33, 48, 61% protonated; neat drug 7%rotary-evaporated powders, XPStrasi2020 R11, C14
lumefantrine with HPMCP, HPMCAS (20–40% drug), CAP, Eudragit L100 (40%)14–42% protonated; amorphous drug 3%powders precipitated into dilute HCl, XPS[bhujbal2021 R8]
fluorescein in miconazole–PVPVA filmslocated in the drug-rich phase, read as a possible acid–base contact with the weakly basic drug ("may arise")confocal dye screen[saboo2017 C36]

Evidence table: acid-base interaction.

Pitfalls and disagreements

The strength of the contact was not measured, and the causal role of the E100 contact in stability was not separated from other properties of E100 wegiel2013a X10. The naringenin–E100 spectra are described but not shown, and no polyphenol-side marker is reported wegiel2013a C17, X10. The melt spectra behind the PSSA reactions are not shown, and no acid–base reaction was seen for bifonazole [vaneerdenbrugh2011a X13, R16]. The lumefantrine values come from dry compacts and fall after dissolution, except for CAP; an earlier XPS study found no protonation with HPMCAS where this one found 19% hiew2022a R20, R30, X14. An ion pair does not guarantee stability: the 50% HPMCAS dispersion, with 1.61 mol drug per mol acid groups, crystallized within a week hiew2022a R26, C63. For curcumin the extent of proton transfer was not measured, and no polymer with the same backbone without amine groups was tested wegiel2014 C37. For resveratrol the IR-based ranking that put E100 first did not hold in storage, where PVP outperformed it wegiel2013b X10. In the lumefantrine powder studies the share of protonated drug ordered the stability of powders made without water but not of powders precipitated into water, where residual water and drying time decided [trasi2020 C36; bhujbal2021 C50, C51]. The polymer that protonated the drug most, CAP, released the least drug at 40% loading [trasi2020 R14; bhujbal2021 R15].

← Concepts and formulas

concept · as used in Crystallization tendency

activity coefficient

The ratio of the ideal solubility to the real solubility of a drug in a solvent, \gamma_d; a value below 1 means that the drug dissolves better than ideal, far above 1 that it mixes much worse [baird2011 E7, C23; marsac2008b E2, C14]. If the entropy of mixing is close to ideal, \gamma < 1 points to exothermic mixing and favourable drug–solvent interactions and \gamma > 1 to endothermic mixing [baird2011 C23; marsac2008a C34]. New aspect (C. Amorphous solubility and LLPS): the activity coefficient γ of the drug in the dilute aqueous solution links concentration to activity, a = C\gamma or a = x\gamma; the amorphous solubility estimate assumes that γ in the bulk is the same at the crystalline and at the amorphous solubility, a "reasonable assumption" for dilute solutions of poorly soluble compounds ilevbare2013a E7, C55; indulkar2015 C43; mosqueragiraldo2015 E5. Micelles break the assumption of equal activity coefficients at the crystalline and the amorphous concentration: for atazanavir in 3.6 mg/mL SDS the slope of free against total drug, labelled γ, was 0.047, so the concentration ratio is about half the activity ratio [indulkar2017a R11, E8, C44]. For telaprevir the ratio of flux slopes below the plateau gives the ratio of activity coefficients (Eq. 7) [lu2017b E7]. New aspects (Miscibility and drug-polymer interactions): For a drug in a polymer the activity coefficient γ carries any non-ideal entropy or heat of mixing, and the mole-fraction solubility of the crystal is its ideal value divided by γ marsac2009 E1, C4. It splits into a combinatorial (entropy) part and a residual (enthalpy) part; exothermic mixing lowers γ and endothermic mixing raises it marsac2009 C10. In a polymer γ is much higher than in a small-molecule analogue because the polymer contributes less mixing entropy marsac2006b C50; marsac2009 C33. Estimating the solubility of a drug in a liquid is essentially the problem of determining γ, which is exceedingly difficult for drugs in viscous glassy polymers marsac2009 C4. New aspect (Bioperformance): γ appears in a = \gamma C for the donor and as γm for the membrane in the flux equation, and additives change the donor value [raina2015c E3, E4, C35]. The concentration-based supersaturation ratio assumes \gamma/\gamma_{eq} = 1; this holds in simple buffers, where the dilute solution is nearly ideal, and fails in media with micelles, where the solubilized fraction differs between the saturated and the supersaturated solution [elkhabaz2019b E2, C28, C42]. New aspect (Physical stability): the activity coefficient of water in an amorphous drug, from water uptake by Raoult's law a_{water} = \gamma_{water} X_{water}, and of the drug in water, from aqueous solubility; both were tested as predictors of moisture-induced separation and showed no trend [rumondor2009c E1, E2, C29].

Findings

PEG dispersions: F3, F11. C. Amorphous solubility and LLPS: F5, F12, F16. Miscibility and drug-polymer interactions: F8, F9. Bioperformance: F3, F6, F18. Physical stability: F2, F13.

Related concepts

depends on ideal solubility; measured by drug solubility in polymer; indicates enthalpy of mixing; related to Flory-Huggins interaction parameter and drug-polymer interactions, and chemical potential; part of thermodynamic activity; depends on combinatorial entropy of mixing.

Formulas

activity coefficient from solubility, Flory-Huggins melting point depression, activity from concentration, binodal condition, solid-liquid equilibrium solubility equation; Flory-Huggins activity equation.

How it is measured or obtained

From the mole-fraction solubility in a liquid analogue of the polymer, PEG 400, after 24 h at 25 °C with a UV assay, and the ideal solubility from the melting point and heat of fusion, \gamma_d = \frac{1}{x_d}\exp\left(\frac{-\Delta H_{fus}(T_m - T)}{R\,T_m^{2}}\right) [baird2011 P5, P9, E7; marsac2008b P5, P9]. PEG 400 is used because activity coefficients in viscous, semicrystalline PEG 3350 are hard to determine [baird2011 C21]. In the amorphous solubility papers it is not measured; in the liquid–liquid equilibrium form x_{L_1}\gamma_{L_1} = x_{L_2}\gamma_{L_2} it links the water-phase and drug-rich-phase compositions ilevbare2013a E12. For the PVP dispersions, from the solubility of the crystalline drug in 1-ethyl-2-pyrrolidone (excess drug, 25 °C, at least 24 h, UV–Vis, or HPLC with refractive-index detection for sucrose) and the ideal solubility from the melting point, heat of fusion and heat capacities; Eq. 8 then scales γ to PVP, assuming ideal entropy with EP, equal contacts and equal γ at both solubility limits marsac2009 P2, P7, E1, E8, C11. In the Bioperformance papers it was not measured; the slope of free against total drug is taken from atazanavir–SDS work [indulkar2017a R11, E8], and the solubility split into a crystal term and an activity-coefficient term was used to compare ritonavir and atazanavir [alhalaweh2016 E3, E4, E5]. In the moisture study, the water activity coefficient from the uptake of the amorphous drug at 95% RH and the drug activity coefficient from mole-fraction solubility [rumondor2009c P13].

Values in the library
materialvalue or classconditionspaper ids
chlorpropamide; aceclofenac; loratadine; haloperidol in PEG 4000.614; 0.121; 2.369; 0.84225 °C[baird2011 R9.a, R9.b, R9.c, R9.d]
acetaminophen, ketoprofen, naproxen, ibuprofen, felodipine, ketoconazole in PEG 4000.06, 0.31, 0.02, 0.64, 0.14, 0.0925 °Cmarsac2008b R14.a, R14.b, R14.c, R14.d, R14.e, R14.f
the same six drugs in water1.2 × 10¹, 1.4 × 10⁴, 3.8 × 10³, 3.1 × 10⁴, 2.4 × 10⁵, 5.3 × 10⁵cited aqueous solubilitiesmarsac2008b R14
nifedipine; felodipine in water1.89 × 10³; 2.43 × 10⁵25 °Cmarsac2008a R10.c, R10.d
five compounds in an ethyl-pyrrolidone analogue of PVPbelow 1 for all25 °Cmarsac2009 C23, R9
felodipine, nifedipine, ketoconazole, sucrose, indomethacin in EP0.89, 0.26, 0.68, 0.37, 0.1525 °Cmarsac2009 R9
same five in PVP K123.94, 1.18, 5.04, 1.76, 0.50scaled with Eq. 8marsac2009 R9
same five in PVP K29/324.34, 1.30, 5.92, 1.93, 0.54scaled with Eq. 8marsac2009 R9
nifedipine, felodipineχ-based activity from melting depression; no direct γFlory–Hugginsmarsac2006b E6
ritonavir, atazanavirlog γ ordering RTV > ATV (recomputed 4.65 against 3.01)pH 6.8[alhalaweh2016 X1, X2]
felodipine / nifedipine fluxγ/γ* assumed 1 for dilute solutionsbuffer[raina2015c C32]
atazanavir, posaconazoleconcentration-based and activity-based ratios agree in the two buffers and differ in the four micellar mediasix media, 37 °C[elkhabaz2019b R8, R9, C42]
water in nifedipine, felodipine, droperidol, pimozide, indomethacin, ketoprofen3.19, 4.13, 3.00, 4.65, 2.54, 3.3525 °C, 95% RH[rumondor2009c R24]
same drugs in water4.95 × 10⁵ to 8.88 × 10⁶; ketoprofen 9.85 × 10⁴from solubility[rumondor2009c R24]

The amorphous solubility chapter prints no activity coefficient value.

Pitfalls and disagreements

The conversion from mg/mL to mole fraction is not printed in Baird and Taylor; the printed values equal moles of drug per mole of PEG 400 in 1 mL, and a true mole fraction would give 0.766, 0.149, 2.49 and 0.856 instead of 0.614, 0.121, 2.369 and 0.842, leaving the split at 1 unchanged [baird2011 X4]. Loratadine, grouped with haloperidol as close to ideal, dissolves at about 0.42 of its ideal solubility [baird2011 X13]. Several rows of the Marsac and co-workers table are not internally consistent, aqueous solubilities come from three cited sources at unstated temperatures and the ideal solubility temperature is not stated, so the coefficients in water serve only as orders of magnitude marsac2008b X5, X18. Activity coefficients in PEG 400 were measured at 25 °C, far below the 52–61 °C of the molten dispersions, and are not values for molten PEG 3350 [baird2011 X16]. A value below 1 in PEG 400 did not separate the crystalline from the transforming drugs in PEO under humidity: all six were below 1 marsac2008b R14, R16. Ketoconazole has 0.09 in PEG 400 but a positive interaction parameter in PEG 3350 marsac2008b R14.f; zhu2011 R5.f. In the amorphous solubility papers the constancy of γ underlies the claim that a solubilizer leaves the maximum supersaturation unchanged, while a later study found the concentration ratio falling [ilevbare2013a C54; raina2015c C29]. There γ is estimated with the ideal dilute solution as standard state, from a buffer calibration [indulkar2017a R11], and is the factor in a = C\gamma; the PEG dispersions chapter defines \gamma_d instead as the ratio of ideal to real solubility in a liquid polymer analogue, so the values tabulated above are not those of the aqueous solutions. The predicted γ in PVP exceeds 1 for four of five compounds (1.18–5.97), so the compounds that dissolve better than ideal in the analogue are predicted to dissolve worse than ideal in the polymer; only indomethacin stays below 1 marsac2009 R9. Lattice volume, heat capacities, ideal solubilities and the mole-fraction basis in PVP are not printed marsac2009 X4, X13. The value of indomethacin in K29/32 differs between table and text marsac2009 X2. In the Bioperformance papers γ/γ* = 1 breaks down with solubilizers [raina2015c C36]; the plotted log γ in the ritonavir–atazanavir figure does not match the recomputation [alhalaweh2016 X1, X2]. The value of \gamma/\gamma_{eq} is not measured; its departure from 1 is read from the gap between the flux ratio and the concentration ratio (F18) [elkhabaz2019b C21, C28]. In the 2009 moisture study the drug-in-water column of the table equals the inverse mole-fraction solubility rather than the printed equation, so it ranks drugs by solubility alone [rumondor2009c X1]; two text rankings disagree with the table [rumondor2009c X2, X3].

← Concepts and formulas

concept · as used in Crystallization tendency

amorphous-amorphous phase separation

The separation of an amorphous drug–polymer mixture into two amorphous phases of different composition, a drug-rich and a polymer-rich one, shown by two glass transitions or by imaging. In the miscibility papers it is the outcome of a miscibility test: solvent-cast films of itraconazole–HPMC (10–50% drug) and telaprevir with HPMC, PVPVA or HPMCAS carried discrete drug-rich domains in a polymer-rich matrix, and blends of long dextran and PVP chains split into a PVP-rich and a dextran-rich phase [purohit2015a C36, C37; li2016c C54; vaneerdenbrugh2012a C16, C17]. Both phases usually hold some of the other component, by partial miscibility or spectral mixing [li2016c C14]. The papers expect the drug-rich phase to crystallize faster, because less polymer surrounds the drug, and a polymer-rich phase to dissolve first [purohit2015a C51; li2016c C41]. Outside the subcategory it is produced by humidity (felodipine–PVP, miconazole–PVPVA, evacetrapib–HPMC) and by water or alcohol in the casting solvent (itraconazole–HPMC) [marsac2010 C14, C27; saboo2017 C42; li2017 C32; mugheirbi2017 C39; mugheirbi2018 R1]. New aspects (ASD process): it arises either because the pair is not miscible at the composition and temperature (thermodynamic) or because the solvent path through a miscibility gap, or later water, traps a separated state (kinetic) [mugheirbi2017 C3; li2017 C1]. The drug-rich phase can crystallize more readily and phase separation can lower the maximum concentration on dissolution [li2017 C1]. In spray-dried ritonavir–PVPVA powders it is read by aged DSC, a pyrene probe and XPS, and it lowered the 25% DL dissolution rate where the baseline release was fast [li2020 R4, R6, R7, R10]. In hydrating ritonavir–PVPVA tablets the drug-rich phase appears as small discrete droplets in a polymer- and water-rich gel (uncoated) or as connected drug-rich regions along particle boundaries (coated), imaged by confocal fluorescence microscopy [benson2025 R14, R16]. New aspect (Physical stability): the moisture-induced form in drug–PVP dispersions, its signatures (IR toward a non-interacting physical mixture, a second Tg near the drug's Tg, near-additive water uptake) and its consequence: crystallization then proceeds from a polymer-poor drug-rich phase [rumondor2009c C15, C24, C5, C6; marsac2010 C49]. New aspect (Release mechanism and LoC): water-induced AAPS of the hydrating dispersion is the authors' explanation of the limit of congruency. Two roles were proposed. As a source of nanodroplets (mechanism II, cited), drug-rich domains formed in the hydrated matrix would be released as droplets; the 2017 nifedipine and ritonavir papers found no evidence for this, the matrix domains being micron-sized against droplets an order of magnitude smaller [indulkar2017b C7, C26; purohit2017b C56]. As a cause of incongruent release, AAPS at the surface leaves a drug-rich layer once the polymer-rich domains dissolve; films above the limit separated on contact with water and films below it did not [indulkar2019a C8, C33; que2019 C35; saboo2020a C43]. If a drug-rich layer forms by AAPS, the drug controls release and the carrier effect of the polymer is lost [indulkar2017b C33]. Papers of 2020–2022 distinguish the morphology of the separated phase: discrete drug-rich domains went with congruent release and contiguous ones with incongruent release [saboo2021 C30, C53; correasoto2022a C53], while release of two high-Tg drug forms failed without visible separation [correasoto2022b R9; hiew2022b R19]. In dissolving compacts the high water content of the gel layer makes water-induced separation possible, favoured close to the solution where water is highest [yang2022a C44].

Findings

Miscibility and drug-polymer interactions: F18, F19, F22, F3, F23, F26. ASD process: F1, F6, F7, F14, F3, F20, F21, F48. Physical stability: F7, F12, F13, F14, F15, F23. Release mechanism and LoC: F5, F8, F12, F20, F22, F25, F27.

Related concepts

contrasts with drug-polymer miscibility; characterized by phase-separated domain size and spectral mixing; persists through kinetic trapping; lowers physical stability. Caused by moisture-induced phase separation or solvent-induced phase separation. Reversed by remixing of separated amorphous phases; driven in humid storage by the Δχ effect; set by drug-polymer-water miscibility. Its rate against dissolution is phase separation kinetics; produces surface drug enrichment; favoured by hygroscopicity; its morphology is phase separation morphology.

Formulas

Gibbs free energy of mixing, additive moisture sorption, multicomponent Flory-Huggins free energy of mixing, Δχ effect.

How it is measured or obtained

Two Tg events by DSC (one Tg is read as miscible, with the limits in single-Tg miscibility criterion) [vaneerdenbrugh2012a P3, C3]; pyrene and prodan fluorescence and fluorescence imaging, AFM-nanoIR, nanoTA, Lorentz contact resonance and AFM topography [purohit2015a P3, P4, P9, P10; li2016c P2, P4, P7, P8]; for polymer pairs also the hydrogen-bonded fraction of PVP carbonyls by FTIR [vaneerdenbrugh2012a P5]. In the later process papers: two Tgs by DSC, AFM with LCR or nanoTA, fluorescence microscopy and the drug's autofluorescence, TEM with EDX or HAADF [li2017 P9, P10, P13; mugheirbi2017 P8, P9; li2019 P10]. In the moisture papers: IR against a physical-mixture spectrum built from C–C band ratios; DSC wet and re-dried; sorption against additivity; AFM and TEM [rumondor2009c P7, E7, P9, P12; marsac2010 P14, P15]. In the release-mechanism papers: Pyrene I/III ratio, nile red or prodan confocal fluorescence, AFM and DSC of films and powders after 97–100% RH, or of films dipped in buffer for seconds [purohit2017b R2, R3, R4, R5, R6; indulkar2019a R18; que2019 R14; saboo2019 R19; saboo2020a R23]; pyrene and TEM of spray-dried particles [bhardwaj2018 R15, R16].

Values in the library
materialvalue or classconditionspaper ids
itraconazole–HPMC 10:90 to 50:50drug-rich domains at every loading by four film methods; one broad Tg by DSC of powderssolvent-cast from methanol–dichloromethane[purohit2015a R1, R11, R13, R7]
telaprevir–HPMC, –PVPVAdomains from 10% drugspin-coated films[li2016c R4, R6]
telaprevir–HPMCASregular domains only from about 30–40% drugspin-coated films, humidity not controlled[li2016c R5, C17, X2, X9]
dextran–PVP, 20 pairsphase separated at 0, 0–5, 5–7, 5–9 of nine compositions for DEX1, DEX6, DEX40, DEX450DSC, freeze-dried[vaneerdenbrugh2012a R13]
felodipine–PVP, 30–50% PVPdrug-rich and polymer-rich phases after 94% RHIR, TEMmarsac2010 C14, C27
miconazole–PVPVA 50:50domains after film formation at 60% RH and after 75% RH storageAFM, confocal[saboo2017 R13, R15, C42]
telaprevir–PVPVA, 70% PVPVAtwo overlapping Tgs; "may have partially phase separated"mDSC[mosqueragiraldo2018b R23, C53]
ECP–polymerdrug-rich spheres (preparation) or drug-rich network (storage)TEM-EDX[li2017 R18, R19, R22]
ITZ–HPMCdiscrete drug-rich domains with self-associated ITZ (extra emission near 381 nm)FLOM, AFM, UV, emission[mugheirbi2017 R1, R22]
LPV–HPMCdiscrete drug-rich domains 50–500 nm high in a polymer-rich matrix; faster release at 33–85% DLAFM, nanoTA, release[li2019 R8, R2]
ritonavir–PVPVA spray-driedsecond non-reversing DSC event after ageing in 50% DL powders from 10:90 to 40:60; drug-rich surfaces (XPS) from 60:40; SNDR at 25% DL about 3.5 → 2.27 → 0.90 mg·min⁻¹·cm⁻¹BLD-35 dryer[li2020 R4, R7, R10]
pimozide–PVP 50:50Tgs −21/44 °C wet, 61/112 °C re-dried94% RH, 42 h[rumondor2009c R17]
felodipine–PVP 46.5 wt%event at ≈45–50 °C instead of one Tg near 80 °Cafter 75–94% RHmarsac2010 R14
felodipine–PVP 50% filmislands of about 300–500 nm (TEM, AFM)94% RHmarsac2010 C41
felodipine–PVP at 93% RHcrystallization independent of PVP level; plateau 70–80%powders[rumondor2009b R21, R11, C20]
felodipine, quinidine, pimozide with PVPVA; pimozide with HPMCAStwo Tgs after 94% RH50:50 powders, dried[rumondor2010c R3.a, R3.c, R3.d, R3.e]
indoprofen, clofoctol, probucol, nilutamide, loratadine with PVPseparation at 70–90% PVP for all; at every composition for nilutamide and loratadineIR, 54–94% RH[rumondor2011 R14]
ritonavir with PVP, PVPVA; HPMCASall PVP and PVPVA dispersions separated, from about 1 h; HPMCAS stayed miscible97% RH, films and powders[purohit2017b C1, C22]
ritonavir 50:50 PVP filmdark domains of about 1–3 µm in a drug-rich continuous phase after 10 s in buffer; 10:90 uniformdried after immersion[purohit2017b R4]
ritonavir–PVPVA films 20, 30, 40%uniform; sparse bright domains; dense domains (LoC 25%)seconds in water[indulkar2019a R18, C20]
ledipasvir–PVPVA films 5, 10, 50%mostly homogeneous; small domains; ≈10–20 µm domains100% RH, 37 °C, 3 days[que2019 R14, C16]
nilvadipine 10:90 film and compactdrug-rich circles and rings of roughly 5–15 µm; rough surface gone after 15 min of dissolution ("superficial")97% RH, 12 h[saboo2019 R19, R17, C24]
felodipine 30% films with PVP, PVPVA, HPMC; HPMCAS, EUDSdiscrete domains; uniform97% RH, 24 h[saboo2020a R23, C23]
ITZ–PVPVA 20% spray-dried particlesno separate drug-rich and polymer-rich phases by pyrene or TEM, also with water in the solventbefore dissolution[bhardwaj2018 C35, C37]
felodipine 30% films with 0–10% TPGSall separated after 12 h; semi-continuous (0, 2%) against discrete (5, 10%) drug-rich domains≈100% RH[saboo2021 R21]
clopidogrel 40, 50, 60% filmsuniform; bicontinuous; drug-continuous5 min, 100% RH[correasoto2022a R20]
atazanavir 10% filmuniform for 3 h at 37 °C; separated within 10 min at 50 °C100% RH[correasoto2022b R9, R12]
lumefantrine free base, benzoate; sulfate filmsseparated; homogeneoushigh RH, 37 °Chiew2022b R19
ritonavir–PVPVA 30:70:5 with Tween 80; Span 20Tg onset of surface material 66.01 → 40.25 and 67.85 → 42.18 °C after dissolution, read as a ritonavir-rich phaseDSC, scraped after dissolutionindulkar2022 R19, C19
ritonavir–PVPVA 10–40% compactsdroplets near the gel–solution boundary (10–20%); continuous drug-rich layer at the interface (30–40%)real-time confocal, pH 6.8[yang2022a R10, R11, R12, R13]
MePHPH–PVPVA; PHPH–PVPVA compactsseparation in the gel layer of most compacts; segregative (MePHPH) and associative (PHPH)in situ confocal with two dyes[deac2023a R6, R7, C25, C28, C71]

Evidence table: amorphous-amorphous phase separation.

Pitfalls and disagreements

A single DSC Tg did not show the separation of itraconazole–HPMC, which the authors attribute to mixing during the scan, a reading supported only by the narrower second-run steps [purohit2015a C39, C26, X1]. Whether the separation is kinetic (solvent-induced, water-induced) or thermodynamic is not settled, and the phase compositions were not measured: the band-ratio and softening data rank the phases but give no weight fractions [purohit2015a C48; li2016c X3, X10]. The separation depended on preparation: humidity, solvent water and the alcohol of the solvent changed whether films separated [mugheirbi2017 C39; mugheirbi2018 R1, C50]. For polymer pairs the plot of the miscibility map is not monotonic cell by cell [vaneerdenbrugh2012a X10]. Separation during dissolution or storage was invoked for PVPVA dispersions of lumefantrine but not measured hiew2022a C43, C61, X11. DSC and polarised light microscopy may overestimate miscibility [mugheirbi2018 C4]. Whether separation helps or hurts release depends on the resulting microstructure [li2019 C5, C58]. The same separation can lower release (ritonavir–PVPVA at 25% DL) or raise it (lopinavir–HPMC at 33–85% DL) [li2020 C64; li2019 R2]. In the humid felodipine–PVP films the composition of the polymer-rich phase was not determined either; the DSC "second Tg" near 124 °C may be depressed melting marsac2010 X7, C18. Faster crystallization after moisture-induced separation was shown across different drugs, not for one drug with and without separation [rumondor2009c X7]. In the 2019 ritonavir–PVPVA study AAPS was seen only in films dipped for seconds or in humidified samples, not in a dissolving tablet, and neither its rate nor that of dissolution was measured [indulkar2019a X6]. A drug-rich porous surface also follows from faster polymer dissolution alone [saboo2020a X6]. In the 2022 ritonavir surfactant study separation was inferred from one lowered Tg of material scraped from the wet tablet, without a second Tg or water content, and was not tested for three of the five surfactants indulkar2022 X10, X4.

← Concepts and formulas

concept · as used in Crystallization tendency

amorphous-crystalline free energy difference

The Gibbs free energy of converting the crystal to the amorphous drug (supercooled liquid or glass) at the test temperature; it fixes the factor e^{\Delta G/RT} of the amorphous solubility almeidaesousa2015 E1, E11. It is estimated from Tm and ΔHf by the Hoffman equation or from temperature-dependent heat capacities by a thermodynamic cycle (the Cp method), to the supercooled liquid or, below Tg, to the glass almeidaesousa2015 E2, E9, E10, E14.

Findings

F5, F7.

Related concepts

estimated by the Hoffman equation; depends on degree of supercooling; sets the amorphous solubility advantage; reference state discussed under structural relaxation.

Formulas

Hoffman equation, amorphous-crystalline free energy difference from heat capacity, Gibbs free energy relation.

How it is measured or obtained

DSC melting data and modulated-DSC reversing heat capacities fitted linearly in T for crystal, glass and supercooled liquid; the Cp method needs an un-ionized, non-solvated crystal that melts without degradation almeidaesousa2015 P5, P6, C11; mosqueragiraldo2015 P3.

Values in the library
materialvalue or classconditionspaper ids
ibuprofen, phenylbutazone, tolnaftate3.24/3.56; 4.84/4.59; 5.69/6.13 kJ/mol (Hoffman / Cp, to supercooled liquid)25 °Calmeidaesousa2015 R11
diclofenac; sorafenib; griseofulvin8.51/9.99; 10.33/10.34; 9.69/9.39 kJ/mol25 °Calmeidaesousa2015 R11
sorafenib; griseofulvin, to the glass11.37; 10.38 kJ/mol25 °Calmeidaesousa2015 R12
CRH15.68 (Hoffman); 5.85 (Cp, supercooled liquid); 5.95 kJ/mol (Cp, glass)25 °C[almeidaesousa2016 M1]
telaprevir, Hoffman13.31 kJ/mol (extractor recomputation)37 °Cmosqueragiraldo2015 E1
atazanavir14.55 kJ/mol, Hoffman, recomputed from the printed Table I inputs (ratio about 62, as printed, with 0.22 read as exp[−I(a)]; about 228 read as I(a))310 K[indulkar2017a E2, X1; indulkar2015 M2]
Pitfalls and disagreements

The Cp parameters are not printed, so the Cp values cannot be recomputed almeidaesousa2015 E9, E10. The claim that the Cp method is more accurate presupposes that the onset equals the supercooled-liquid solubility almeidaesousa2015 X4. The glass carries more free energy than the supercooled liquid below Tg almeidaesousa2015 C29. For atazanavir (undercooling 171 K, synthesis: 481 − 310 K) the Hoffman inputs printed give the printed ratio of 62 only with 0.22 read as exp[−I(a)], not I(a) as Table I heads the column (synthesis) [indulkar2017a X1; indulkar2015 M2]; for pazopanib the cycle cannot be run because it degrades on melting [sugihara2018 C17].

← Concepts and formulas

concept · as used in Crystallization tendency

amorphous salt

The amorphous form of a drug salt; salt formation adds an ionic interaction between the protonated base and the counterion, taken to dominate over hydrogen bonding in setting Tg towler2008 C23. Combining salt formation and amorphization "may be desirable", for example for a lyophilized product towler2008 C18.

Findings

F20.

Related concepts

depends on glass transition temperature; related to crystallization tendency and physical stability.

Formulas

none in this chapter (counterion descriptors listed in the chapter table).

How it is measured or obtained

Prepared by freeze-drying or melt quenching and checked by DSC towler2008 P3, P5, R1; Tg, recrystallization on heating, moisture sorption and storage at high humidity are then measured towler2008 P6, P8, P9; later by open-dish storage at 40 °C/75% RH hiew2022b R7; duong2022a C32.

Values in the library
materialvalue or classconditionspaper ids
propranolol saltsTg 9.0–35.9 °C against −9.3 °C for the baseDSCtowler2008 R4
nicardipine saltsTg 28.5–81.0 °C against 3.2 °C for the baseDSCtowler2008 R5
20 salts of both bases11 amorphous after freeze-dryingDSCtowler2008 R1
propranolol salts0.39–61.90% water uptake at 75% RHvapour sorptiontowler2008 R8
lumefantrine saltsbenzoate, PEG 250 diacid, besylate crystallized within 4 weeks; HCl and sulfate not for at least 16 weeks40 °C/75% RHhiew2022b R7
delamanid saltsstable 2 weeks; salt formation raised Tg40 °C/75% RHduong2022a C32, C47
Pitfalls and disagreements

Stoichiometry and ionization state of the amorphous salts were not reported towler2008 X8. Crystallization and moisture uptake did not follow counterion pKa towler2008 C14, C34; drug or salt Tg appeared an important predictor for lumefantrine hiew2022b C37, yet the highest-Tg propranolol salt crystallized towler2008 R12. Salt formation did not change delamanid crystallization in dispersion films at pH 3.0 nguyen2023b C17.

← Concepts and formulas

concept · as used in Crystallization tendency

amorphous solid dispersion

A drug dispersed in a polymer in the amorphous state to raise dissolution rate and supersaturation. In the amorphous solubility and LLPS papers an ASD matters as a route to supersaturation: dissolved beyond the amorphous solubility it gives a drug-rich phase and a cap on the free drug (F8, F13) [ilevbare2013a C52, C5; almeidaesousa2016 C6]. The ASDs tested were ritonavir:PVP 1:9 and CRH1:HPMC 20:80 [ilevbare2013a P13; almeidaesousa2016 M7]. The ueda2020b, ueda2021 and ueda2022 studies add that the polymer of an ASD, once dissolved, enters the drug-rich phase and lowers the amorphous solubility, so a higher polymer:drug ratio in the ASD can "conceivably" reduce the supersaturation advantage further; the studies used dissolved polymers, melt-made ibuprofen ASDs for DVS and solid-state NMR, and no dissolving ASD [ueda2020b C37, C55; ueda2021 P6; ueda2022 P5]. New aspects (Miscibility and drug-polymer interactions): A molecularly mixed, single-phase, homogeneous mixture of drug and polymer, in older wording a "glass solution" in which a solute is dissolved in a glassy solvent kestur2010b C4; taylor1997 C1. Amorphous dispersions are used to raise apparent solubility and dissolution, but the amorphous phase is metastable and may crystallize during storage or dissolution taylor1997 C1, C2; vaneerdenbrugh2010b C6, C7. The miscibility papers hold that the polymer must be in the same amorphous phase as the drug to be effective ("it is generally accepted") and that maximum stabilization "can only be realized" with intimate mixing, so that a physical mix does not inhibit crystallization [kestur2010a C2; rumondor2009a C3]. Dispersions made by solvent evaporation were not always molecularly mixed: itraconazole–HPMC and telaprevir films carried drug-rich domains [purohit2015a C36; li2016c C54], and neat naringenin and quercetin could not be made amorphous by rotary evaporation, so a polymer was needed to make the dispersion at all wegiel2013a R7, C46. Ionizable polymers (E100, HPMCAS, HPMCP, CAP, Eudragit L 100, PAA) add acid–base contacts wegiel2013a M4, M8; hiew2022a M3, M4, M5, M6. New aspects (ASD process): the earliest paper of that chapter asks which carrier and process give a useful solid form of an extremely poorly soluble drug and finds that the carrier set whether probucol stayed amorphous [broman2001 C1, C14]. (synthesis: the paper's "solid dispersions" include samples that are not amorphous, since the probucol–PEO and probucol–PAA dispersions are mainly crystalline Form II [broman2001 R4.c–f].) The ASDs of the process papers were also made by solvent granulation of drug and polymer onto lactose and cellulose trasi2019 C50, by spray drying from water-containing methanol [li2020 P1] and by extrusion of physical mixtures of drug and polymer moseson2020 P4. New aspect (Bioperformance): the ASD is a drug dispersed amorphously in a polymer (or, in alhalaweh2016, in a mix with a second drug) that releases drug into supersaturation. Rotary-evaporated dispersions were used in alhalaweh2016, hate2018, mosqueragiraldo2018b and elkhabaz2017, and commercial capsules in purohit2018 [alhalaweh2016 P3; hate2018 P5; mosqueragiraldo2018b P1; elkhabaz2017 P6; purohit2018 M2]. Later papers dose such dispersions to animals: rotary-evaporated enzalutamide dispersions to rats, spray-dried pretomanid tablets to monkeys, and generic and innovator tacrolimus capsules to dogs, the generic being a mainly physical mixture of amorphous drug with HPMC and the innovator an intimate dispersion [wilson2018 P10; wilson2020 P1; nguyen2023a P4, P5; purohit2019 C21]. New aspect (Physical stability): the physical-stability papers call the drug–polymer glass a molecular (one-phase) dispersion when it shows a single Tg and drug–polymer hydrogen bonds by IR [konno2006 C8, R5; rumondor2009c C1], and stress that a polymer can inhibit crystallization only while it stays intimately mixed with the drug marsac2010 C31. In the study of marketed tacrolimus capsules the term is used strictly: a product that contains the polymer but holds the drug as an amorphous coating on lactose, not mixed with HPMC, is in the authors' belief not an amorphous solid dispersion, and molecular-level mixing is "considered to be essential" for stabilization trasi2017 C47, C50.

Findings

C. Amorphous solubility and LLPS: F8, F13, F14, F19, F20, F21. Miscibility and drug-polymer interactions: F1, F3, F9, F15, F16, F18, F19, F23, F26. ASD process: F17, F18, F19, F22, F23, F24. Bioperformance: F6, F8, F9, F11, F16, F23, F24. Physical stability: F1, F12, F14, F23, F24, F26, F27, F28, F29, F30, F33, F35.

Related concepts

produces a drug-rich phase above the amorphous solubility; relieved by crystallization unless a polymer is present (F14); requires drug-polymer miscibility; stabilized by drug-polymer interactions and kinetic stabilization; related to crystallization inhibition and supercooled liquid. Processed in the HME operating design space. Sets drug loading effects; related to non-sink dissolution. Can undergo amorphous-amorphous phase separation; stabilized by crystallization inhibition by polymers.

Formulas

Gordon-Taylor equation.

How it is measured or obtained

Rotary evaporation from methanol (ritonavir) or methanol/dichloromethane (CRH1) and incremental or single-dose non-sink dissolution [ilevbare2013a P13, P14; almeidaesousa2016 P2, P11]. The dispersions of the miscibility papers were made by dissolving drug and polymer in a common solvent and removing it: rotary evaporation from methanol, ethanol, chloroform or dichloromethane–ethanol, followed by vacuum drying [taylor1997 P3; rumondor2009a P1; vaneerdenbrugh2010b P6]; spin coating on coverslips, ZnSe or KRS5 for films vaneerdenbrugh2010b P1; marsac2009 P4; cryomilling of a 10% w/w dispersion with crystalline drug and melt-quenching for the low-polymer growth samples kestur2010a P1, P2. A physical mix of amorphous drug and polymer is the control without molecular mixing taylor1997 C24, C26. Amorphous state is checked by XRPD or polarized light taylor1997 R1; vaneerdenbrugh2010b P2. Further preparations: rotary evaporation at 45–60 °C with 24 h vacuum drying and cryomilling wegiel2013a P1; hiew2022a P5; spin coating on KRS-5, ZnSe or quartz [purohit2015a P2, P9]; freeze-drying of aqueous polymer blends [vaneerdenbrugh2012a P2]; 50 mg compacts for release hiew2022a P9. In the process papers the solid form was read by Raman and DSC (single Tg between those of the components; polymer-free marker band) [broman2001 P1, P2, R1, R4]. In the Bioperformance papers the dispersions were made by rotary evaporation from methanol and cryomilling, and checked by PXRD, DSC, PLM and SHG [hate2018 P5, P10; elkhabaz2017 R8]. In the physical-stability papers: spin-coated films (dichloromethane:ethanol, ethanol or chloroform, heated to 90 °C) for nucleation, growth and IR; rotary-evaporated and, for low polymer contents, cryomilled powders for DSC, sorption and PXRD [konno2006 P1, P3; rumondor2009c P1; rumondor2009b P1; kestur2010b P1]. Spin-coated films and rotary-evaporated powders of nine 50:50 drug–polymer pairs [rumondor2010c P1, P5]; rotary evaporation in 5% steps to find the highest amorphous loading wegiel2013b P1; rotary evaporation followed by cryomilling wegiel2014 P2, P3; a 10% PVP dispersion diluted with crystalline drug by cryomilling, then melted, to reach 3% PVP kestur2013a P4; kestur2013b P2.

Values in the library
materialvalue or classconditionspaper ids
ritonavir:PVP 1:96.4, 11.8, 18.3, 18.0 µg/mL with 6.0, 12.0, 18.0, 24.0 added100 mM pH 6.8, 37 °Cilevbare2013a R17
CRH1:HPMC 20:80about 12.9 µg/mL without polymer in the medium; HPLC maximum about 13.2pH 4.6, 25 °C[almeidaesousa2016 R8, R12]
indomethacin–PVP-K90, 5–70 wt% PVPamorphous by XRPD; interactions in dispersions, none in physical mixesmethanol, rotary evaporationtaylor1997 R1, R10, R11, R12
felodipine–PVP, nifedipine–PVPone amorphous phaseIR, PXRD[rumondor2009a C38]
8 drugs × 7 polymers392 spin-coated compositions scored over 7 daysethanol or ethanol–dichloromethanevaneerdenbrugh2010b R2
50/50 drug–PAA, drug–PVP powderscrystalline with PAA for acidic drugs, amorphous with PVProtary evaporation, 7 daysvaneerdenbrugh2010b R9
felodipine, 0.5–4.5% w/w polymermade by diluting a 10% dispersion by cryomillingmelt-quenched filmskestur2010a P1, C5
naringenin, quercetin with six polymershighest X-ray amorphous polyphenol content about 20–85%; E100 highest, PAA lowestrotary evaporation at 60 °C, cryo-milledwegiel2013a R8
lumefantrine with PVPVA and four enteric polymersall X-ray amorphous with one Tg, except 50% HPMCAS (melted at 126.3 °C on heating)rotary evaporation at 45 °C, sieved below 250 µmhiew2022a R4, R6
itraconazole–HPMC, telaprevir–polymersdrug-rich domains in the filmsspin-coated from methanol–dichloromethane[purohit2015a R11; li2016c R4, R5, R6]
probucol–PVP K30 50:50amorphous; Tg 95 °C (solvent), 129 °C (moulded)Raman, DSC[broman2001 R4.a, R4.b]
probucol–PAA, –PEOmainly Form IIsame[broman2001 R4.c–f]
probucol–PVP–PAA 50:25:25mainly amorphousboth routes[broman2001 R4.g, R4.h]
ritonavir and lopinavir with PVPVA 1:1 on lactose:MCCX-ray amorphous granules, unchanged for one month at 40 °C/75% RHsolvent granulationtrasi2019 R1, R2
ritonavir–PVPVA spray-dried, 25–75% DLX-ray amorphous (methanol powders); one Tg of 91, 75 and 63 °Cspray drying[li2020 R1, R2]
ritonavir/atazanavir/lopinavir in PVP K29/3290% polymer, Tg 139–160 °C10% drug[alhalaweh2016 R4]
atazanavir in HPMCAS, HPMC, PVPVA10, 30, 50%X-ray amorphous[hate2018 R10]
telaprevir in HPMCAS, HPMC, PVPVA, CA Sub10, 30, 50%one Tg for HPMCAS 50%[mosqueragiraldo2018b R15, R23]
ezetimibe in PAA, PVP, HPMC-AS50:50no crystals by PLM or SHG[elkhabaz2017 R8]
tacrolimus capsulestacrolimus:HPMC 1:1; drug about 4% of fillgeneric[purohit2018 M2, R2]
enzalutamide in PVPVA or HPMCAS-MF10 and 50 wt% drug, rotary-evaporated, amorphous by XRPD and FTIRused within 2 weeks[wilson2018 P10, P11]
enzalutamide in CA Sub or CPHPC-106CA Sub 50%; CPHPC-106 10 and 50% drugrotary evaporation[wilson2020 P1, A3]
atazanavir in AS-LF or AS-HF1:9, spray-drieddissolution and transport[ueda2020a P1, M14]
pretomanid in HPMCAS-HF, MF, LF blends20% drug in the monkey tablets, spray-dried; screening powders rotary-evaporatedwith TPGS or tris in two tabletsnguyen2023a P4, P5
felodipine with PVP, HPMCAS, HPMC 0–100%single Tg at every compositionrotary-evaporated, DSCkonno2006 R5, C8
five drugs with PVPdry dispersions one phase by IR; single Tg for droperidol and pimozidespin-coated and rotary-evaporated[rumondor2009c R3, R5, C1]
felodipine with 0.5–4.5 wt% PVPmade by cryomilling a 10% dispersion with crystalline drug, then meltingto avoid partial crystallization on rotary evaporationkestur2010b P1
resveratrol with E100, PVP, HPMC, HPMCAS, CMCAB, PAAhighest X-ray amorphous loading 60, 45, 40, 35, 35, 15%rotary evaporationwegiel2013b R2
curcumin with five polymers75:25 dispersionsrotary evaporation, cryomillingwegiel2014 P2, P3
tacrolimus–HPMC 1:1 (solvent cast) and six marketed capsulesdispersion amorphous 4 weeks at 40 °C/75% RH; one generic 57–95% crystalline at 4 weeksopen dishtrasi2017 R16, R13
ATP enantiomers with 5% w/w polymermelt-quenched films from solvent-evaporated mixturesgrowth at 40–90 °C[sato2015 P1, P3]
indomethacin with 10% w/w HPMC, HPMCAS or PVPsolvent-evaporated dispersion, then melt-quenchedgrowth from 80 °C to near Tmtian2017 P2
Pitfalls and disagreements

One composition of one polymer each; the medium of the CRH1 ASD runs in Figs. 7 and 8 is unclear [ilevbare2013a X15; almeidaesousa2016 X5]. The preparation route changes the result: films, rotary-evaporated powders and melt-quenched powders of the same pair can differ (PH–PAA) and the three characterization methods of one study used differently prepared samples [vaneerdenbrugh2010b C44; rumondor2009a X4]. Direct rotary evaporation below 5% w/w polymer would let felodipine crystallize, hence the cryomilled dilution, whose uniformity at the molecular level is assumed, not measured kestur2010a C5. A single Tg does not prove a single phase (see single-Tg miscibility criterion). Thermodynamically stable dispersions are hard to achieve; most are supersaturated and kinetically stabilized marsac2009 C34. The amorphous and single-phase state of the halogen-bond dispersions (up to 50 wt% for XPS, 90 wt% for IR) was not verified [que2020 X10]. Neat naringenin gave an amorphous film only by spin coating, quercetin not at all wegiel2013a R7. Amorphous probucol occurred only with PVP present, so carrier and form are confounded [broman2001 X13]; amorphous does not mean fastest release [broman2001 C35]. Two drugs in one dispersion can form a co-amorphous system whose amorphous solubilities are lower than those of the single-drug dispersions trasi2019 C38. In the Bioperformance papers making an ASD does not by itself remove dissolution-rate-limited absorption (high loading) [indulkar2016 C41]. The lowest loading did not always give the best dispersion (10% enzalutamide in HPMCAS crystallized), and the best solution inhibitor of 24 polymers made a dispersion no better than the crystals (F8, F24) [wilson2018 R6, R16; wilson2020 R1, R6]. A dispersion that is one phase when dry can separate under humidity and stay separated after drying [rumondor2009c C2; marsac2010 C53]. Nucleation was measured on films and Tg on powders made differently, so "one phase" is shown on one sample form and assumed for the other konno2006 X12. In the later felodipine growth papers too, below about 5% polymer rotary evaporation was expected to crystallize felodipine partly, hence the two-step dilution kestur2013b C5. Interactions were read on spin-coated films at other compositions than the stored powders wegiel2014 X8; wegiel2013b X12. Equal composition does not mean equal structure: six tacrolimus products with about one part HPMC per part drug differed in stability because of the manufacturing route trasi2017 R1, R6, C47.

← Concepts and formulas

concept · as used in Crystallization tendency

amorphous solubility advantage

The factor by which the amorphous drug is more soluble than the crystal, e^{\Delta G/RT} from the crystal–amorphous free-energy difference, reduced by the activity of the water-saturated amorphous drug ilevbare2013a E1, C60; almeidaesousa2015 E1. In the amorphous solubility and LLPS papers the measured counterpart is the supersaturation ratio at the LLPS onset (F6). The papers of 2017–2022 add measured values of the ceiling ratio of 63.6 (atazanavir, synthesis: 70 ÷ 1.1), 28.5 (telaprevir, synthesis: 148 ÷ 5.2), about 600 (pazopanib) and 2.5–5.9 (ibuprofen, falling with temperature), and that micelles lower the concentration-based advantage (atazanavir 65 → 33) without lowering the activity-based one [indulkar2017a R4, R6, R13; lu2017b R2, R1; sugihara2018 R4; ueda2022 R16]. New aspect (ASD process): it is the supersaturation an ASD can reach if it stays amorphous and dissolves; crystals in the dispersion "directly" lose their share of it ("potency") and may also consume what is generated [moseson2020a C3]. For bicalutamide the maximum advantage over the stable form is 35-fold [moseson2021 R2]. In the pH-shift test of posaconazole the dose was set at about 10 times the amorphous solubility in the intestinal medium (19.1 µg/mL), that is about 191 µg/mL, and 200 µg/mL at full release [shah2024 P11, X9]. New aspect (Bioperformance): the ratio is computed by the Hoffman estimate with water correction or taken as the LLPS onset over the crystalline solubility [raina2014a R7; raina2015c E8]. New aspect (Crystallization from solution): in the solution-crystallization papers the advantage fixes the highest supersaturation tested: the calculated amorphous solubility over the crystalline solubility gives S 15.3–15.4 for ritonavir and about 14.7 for celecoxib, and the "amorphous benchmark" S of about 10 for felodipine [ilevbare2012b R3; ilevbare2013c E1, R2; alonzo2012 C42]. The measured counterpart is the plateau after a pH shift or the onset of phase separation (F2). For six dihydropyridines the estimated ratios of amorphous to crystalline solubility in buffer were 16.1 (cilnidipine), 9.03, 8.42, 23.0, 11.7 and 22.9, not reproducible from the printed inputs (felodipine 5.93 recomputed against 6.68 µg/mL); for danazol the recalculated estimate of 7.8 µg/mL is a ratio of 8.7 against the earlier 19.6 [raina2015b X2; jackson2016 R3, R4]. New aspect (Supersaturation from ASD dissolution): the calculated advantage is the yardstick for what a dissolving amorphous solid or dispersion should reach, and the question is whether a dispersion can exceed that of the pure amorphous drug: by flux it did not for felodipine, and the authors add that thermodynamics gives "no reason to expect" the two to be equal [alonzo2011 C22, C45]. The advantage is "rarely achieved" because the solid or the solution crystallizes [konno2008a C4; alonzo2010 C2]. For six marketed tacrolimus capsules the advantage of about 35-fold was "sustained throughout" a 2 h non-sink test trasi2016 C44, C16; mixing the drug with a polymer that does not dissolve lowers the advantage a dispersion can realize [li2018 C67].

Findings

Amorphous solubility and LLPS: F5, F6, F7, F12, F14, F16, F21. ASD process: F23, F26, F27, F28, F39, F41. Bioperformance: F1, F4, F18. Crystallization from solution: F2, F9, F14, F19, F21, F22, F34. Supersaturation from ASD dissolution: F1, F5, F7, F18, F22, F23, F28, F29.

Related concepts

same as the ceiling supersaturation ratio; built from amorphous-crystalline free energy difference and water sorption correction of amorphous solubility; gives the amorphous solubility. Realised in non-sink dissolution; lost to residual crystallinity and matrix crystallization; lowered by mutual amorphous solubility reduction. Depends on Hoffman equation; estimated with the Gibbs-Duhem equation water correction; depends on crystalline solubility; related to free drug concentration.

Formulas

amorphous solubility advantage with water-sorption correction, amorphous solubility advantage from free energy difference, supersaturation ratio. Amorphous-to-crystalline solubility ratio, Hoffman equation.

How it is measured or obtained

By calculation (Hoffman or heat-capacity free energy, sorption isotherm) and by the onset almeidaesousa2015 P9, P10; also by dissolving an amorphous film ilevbare2013a P4. In the ASD process papers: crystalline solubility by 72 h shake flask; amorphous solubility as the onset of light scattering during solvent-shift addition (liquid–liquid phase separation), in some cases only with a polymer present because the drug crystallised too fast without it [moseson2020a P7, P8; moseson2021 R1, R27]. In the Bioperformance papers it is estimated from melting data or as onset ÷ crystalline solubility [raina2015c P3, P6]. In the crystallization-from-solution papers: calculated with the Hoffman equation and a water-sorption correction; measured by dissolving a spin-coated film (ritonavir), by pH-induced precipitation, and by the onset of phase separation of a solvent-shifted solution [ilevbare2012b P4–P6; hsieh2012 P11; jackson2014 P6, P7]. In the ASD dissolution papers: calculated from melting data with the Hoffman equation and the measured crystalline solubility [konno2008a P7; alonzo2010 P12; alonzo2011 P8]; tested against flux relative to the crystal [alonzo2011 R14].

Values in the library
materialvalue or classconditionspaper ids
ibuprofen to griseofulvinpredicted 3.7/4.2, 7.0/6.4, 9.8/11.7, 28.9/52.5, 48.4/48.6, 34.8/30.9 (Hoffman / Cp method)25 °Calmeidaesousa2015 R13
clotrimazole; atazanavirpredicted about 9.2 and 62; measured about 13 and about 6337 °Cindulkar2015 R3, R10
felodipine, buffer; with 0.04% vitamin E TPGS; in FaSSIFabout 9.2; 4.5; 5.0 (concentration-based)37 °C[raina2015c C29]
tacrolimusabout 35 (49/1.4), about 33–36 by method; other drugs 5–65amorphous solubility by ultracentrifugation, pH 4.5, 37 °Ctrasi2016 R16, R17, X9
IDM, pH 4.5 acetatecrystalline about 8, amorphous about 80 µg/mL (about 10-fold, synthesis); unchanged by 50 µg/mL PVPVA37 °C[moseson2020a R1]
BCL, pH 6.8 phosphateform 1 4.3, form 2 15, amorphous 150 µg/mL; 35-fold37 °C[moseson2021 R1, R2]
RTV and LPV, pH 6.8crystalline 1.8 and 2.9, amorphous 31 and 19 µg/mL10 mM phosphate, 37 °Ctrasi2019 M1, M2
IDM–PVPVA with 25% residual crystalsC720 about 28.8 against about 43.1 µg/mL: the share of the advantage lost with the crystals50 µg/mL total dose[moseson2020a R6]
posaconazole in FaSSGF and FaSSIF V2LLPS concentration (taken as the amorphous solubility) 2326 and 19.1 µg/mL against crystalline 86.5 and 1.2 µg/mL, ratios about 27 and 16 (arithmetic on the cited values)37 °C[shah2024 R11, C14]
ritonavir / atazanavir / lopinavirabout 14 / 83 / 6pH 6.8[alhalaweh2016 R6]
clotrimazole15pH 8[indulkar2016 R3]
atazanavirobserved ratio in SDS about half that in buffer, with unchanged activity[indulkar2017a C64, C45]
atazanavir, posaconazoleconcentration ratio about 55 and 36 in the buffers, about 14 and 5 in pooled human fluid; flux ratio 50–56 and 29–37six media[elkhabaz2019b R8, R9]
atazanavir, darunavir58 and 1.37 from the printed solubilities (printed "almost 82-fold" and "1.30")pH 6.8, 37 °C[alkalla2026 R3, R6, X1]
pretomanid8–10 in buffers37 °Cnguyen2023a R1
tacrolimusabout 33pH 4.5[purohit2019 C30]
enzalutamideabout 14.5 (42/2.9); S "approximately 13" is the theoretical 39/2.9pH 6.5[wilson2018 R1, R3, X2]
felodipineS of about 10 over the crystal (theoretical and measured)benchmark for the supersaturation range S 3–12[alonzo2012 M1, C42]
ritonavir20.6 µg/mL calculated, 19.8 measured; S 15.3–15.4 over 1.3 µg/mL37 °C[ilevbare2012b R1, R2, R3]
efavirenz; celecoxib19.8 and 22.6 µg/mL calculated against 12.3 and 2.5 measured37 °Cilevbare2013c R2
five weak basesratios 22.2, 11.9, 20.4, 54.6, 7.0 without and 12.3, 9.8, 16.7, 15.3, 4.3 with water correction37 °Chsieh2012 R10
glyburide; warfarinabout 60 and 4825 °Chsieh2014 R16
danazol19.6 printed (15.4 recomputed); earlier estimates 13.7 and 26.525 °C[jackson2014 R2, R3, X1]
six dihydropyridinesestimated amorphous solubility 0.5, 6.68, 48, 20, 6.3, 19.0 µg/mLbuffer, Hoffman with water correctionraina2015b R10
danazol7.8 µg/mL with heat-capacity data and the water correction; onset 8 µg/mL25 °C, pH 6.8[jackson2016 R3, R8]
posaconazolemeasured amorphous/crystalline 33.6 at pH 7.0 (32.4–37.3 from pH 2.0 to 7.0); estimate 37.9 printed37 °Cduong2022b R1, R6, R7, X5
felodipineabout 10 times the crystal, about 9.5 µg/mL37 °C, pH 6.8 test fluid[konno2008a R17]
felodipineabout 9.1 µg/mL at 37 °C; 5–6 µg/mL at 25 °CpH 6.8[alonzo2011 M1]
felodipine, flux ratio to the crystal8.5 (pure amorphous), 9.5 (50:50), 10.5 (90:10 HPMC dispersion)dialysis, 37 °C[alonzo2011 R14]
indomethacinabout 21 µg/mL (crystal about 1.2 µg/mL)pH 2.0[alonzo2011 M2]
ritonavir; lopinavir (cited)25 and 17 µg/mL against 1.7 and 2.7 µg/mL crystallinepH 6.8[trasi2015b M1, M2]
lopinavirabout 5.6 (17.4/3.1)pH 3, 25 °C[li2018 E2, R1]
indomethacin with 100 µg/mL HPMC36 µg/mL against 1.61 (γ) and 2.8 µg/mL (α)pH 2.2, 37 °C[xie2017 R3, R1]
paclitaxel31.6 against 0.36 µg/mL, a ratio of about 88pH 6.8, 37 °C, three methods[que2018 R1, R2]
delamanid0.76 against 0.018 µg/mL at pH 6.5, a measured ratio of 42.3 against 31.8 from the Hoffman free energy37 °Cduong2022a R1, R27
praziquantel, racemate and R-enantiomerratios 5.6–7.7 and 4.0–5.8 across nine media; amorphous 1.17–1.74 and 1.20–1.70 mg/mL37 °C[alvarenga2025 R1]
posaconazoleabout 20 against 1.69 µg/mL in FaSSIF; 115.1 µg/mL crystalline in FaSSGF37 °C[elkhabaz2019a R2, R1]
Pitfalls and disagreements

The calculated advantage fails for high-melting drugs when the Hoffman free energy is used (F7) mosqueragiraldo2015 C23, C24. When additives raise crystalline solubility more than the LLPS concentration, the concentration-based ratio falls (felodipine, about 9.2 in buffer to 4.5 in 0.04% vitamin E TPGS and 5.0 in FaSSIF), while the maximum flux stays in the buffer band, so the advantage in activity is retained [raina2015c R9, R11, C29]. In dispersions, a dose below the amorphous solubility (IDM 50 against about 80 µg/mL) can still lose most of the advantage to crystals; a crystal-free bicalutamide dispersion never reached 40 µg/mL against 150 µg/mL [moseson2020a X17; moseson2021 C41]. The amorphous solubility of ritonavir alone is reduced to about 4 µg/mL in a co-amorphous granule trasi2019 R4. In the Bioperformance papers the ratio from concentrations falls with solubilizers although the flux advantage does not [raina2015c C43]; the atazanavir ratio differs between cards because the crystalline solubility differs (F1). In human fluid the flux of a solution at the amorphous solubility was about 30 times that of a crystalline suspension for posaconazole, so the advantage was not lost although the concentration ratio fell to about 5 (F18) [elkhabaz2019b C30, C45]. The printed atazanavir and darunavir ratios do not follow from the printed solubilities [alkalla2026 X1]. With a polymer in the medium the same ratio applied to the polymer-containing solubility overestimates the ceiling (danazol: 43 against an onset of 8–9 µg/mL) [jackson2014 X9]. Celecoxib's measured 2.5 µg/mL is far below the calculated value and is not discussed ilevbare2013b X9. The efavirenz run started at S about 2.3, the ratio of calculated amorphous to crystalline solubility being 2.4 ilevbare2013c X4. The printed estimates for the dihydropyridines differ between media in ways the equation does not allow raina2015b X2. Felodipine dispersion and solution readings of 14.1 and about 11 µg/mL exceed the 9.5 µg/mL estimate of the 2008 paper, measured by UV through a 70-µm filter [konno2008a X5]. The 25 °C indomethacin range (17–25 µg/mL) does not reproduce from its printed inputs (15.8–23.6) alonzo2010 X2. That a dispersion reached the same activity as the pure amorphous drug "may be purely coincidental" [alonzo2011 C48]. The tacrolimus ratio uses an ultracentrifugation value measured with 10 µg/mL HPMC in the buffer trasi2016 X9. For praziquantel the ratio is lower for the enantiomer because its crystal is more soluble, not because its amorphous solubility is lower [alvarenga2025 R1].

← Concepts and formulas

concept · as used in Crystallization tendency

amorphous solubility

The concentration of dissolved drug at which a supersaturated aqueous solution is in apparent, metastable equilibrium with the amorphous drug, more rigorously with the water-saturated supercooled liquid or glass; it is not a true thermodynamic solubility, which is why the authors keep it in quotation marks almeidaesousa2015 C3, C39. In this chapter it is operationally the concentration at which a drug-rich phase first separates (the LLPS or GLPS onset), and the free-drug ceiling above which added drug only makes more drug-rich phase ilevbare2013a C3, C5; mosqueragiraldo2015 C34. It exceeds the crystalline solubility by e^{\Delta G/RT} and is lowered by the water the amorphous drug absorbs ilevbare2013a C60. The papers of 2017–2022 add that the dissolved-drug concentration in the presence of a drug-rich phase can be read directly by NMR (ibuprofen 154 µg/mL in buffer against 151 µg/mL calculated) [ueda2021 R5, R6, C10], and that the value changes when a second component enters the drug-rich phase or the water phase: a dissolved polymer that enters the droplets lowers it (ibuprofen 177.1 → 96.9 and 90.7 µg/mL with 300 µg/mL HPMC and PVP-VA) and a polymer that complexes the drug in water raises it (256.3 µg/mL with EUD-E) [ueda2020b R1]; micelles raise the total concentration at the amorphous solubility but not its activity [indulkar2017a R13]; STCDC and SGCDC lower the activity at which it is reached [lu2017b R18]. New aspect (Bioperformance): the highest free-drug concentration that an amorphous (supercooled-liquid or glassy) drug can produce in a medium; above it a second, drug-rich phase separates and the free concentration stays at this value [raina2014a C24, C27; alhalaweh2016 C1; hate2018 C5]. Several papers treat the concentration at which the second phase appears (the LLPS or glass–liquid onset) as the amorphous solubility [indulkar2016 R2; hate2018 R2; alhalaweh2016 R6]. In ASD work it is the ceiling of the free concentration a dispersion can give, and the concentration at which flux stops rising (F2). Enzalutamide, atazanavir, darunavir, clozapine, ketoconazole, nifedipine and pretomanid all take the UV-extinction onset as the amorphous solubility, with the polymer or medium of the test present [wilson2018 R2, C16; ueda2020a R1; alkalla2026 R3; nguyen2023a R1, P2]. In rats the enzalutamide dispersions that reached the ceiling gave several times the exposure of the crystalline drug (F23) [wilson2018 R16, R18].

Findings

Amorphous solubility and LLPS: F3, F4, F5, F6, F7, F8, F9, F10, F11, F12, F13, F14, F15, F16, F17, F18, F19, F20, F21, F23. Bioperformance: F1, F2, F4, F6, F7, F8, F15, F18, F19, F20, F23, F27, F30.

Related concepts

depends on crystalline solubility and water sorption correction of amorphous solubility; measured as the onset of liquid-liquid phase separation; sets the ceiling of supersaturation; lowered by a second component in the drug-rich phase; gives the amorphous solubility advantage. Papers of 2017–2022: lowered by a dissolved polymer that enters the drug-rich phase (polymer partitioning into the drug-rich phase); not raised in activity terms by micelles (activity-based supersaturation); its upper limit on the activity scale is the maximum achievable supersaturation. Depends on Hoffman equation inputs; related to thermodynamic activity.

Formulas

amorphous solubility advantage with water-sorption correction, amorphous solubility advantage from free energy difference, Hoffman equation, amorphous-crystalline free energy difference from heat capacity. Amorphous-to-crystalline solubility ratio, ideal-mixing reduction of amorphous solubility.

How it is measured or obtained

(1) As the onset of scattering during slow addition of a drug stock, confirmed by a polarity probe ilevbare2013a P5; almeidaesousa2015 P1, P2. (2) As the drug concentration in the supernatant after ultracentrifugation of a phase-separated solution [trasi2015a P2; almeidaesousa2016 P7]. (3) By dissolving an amorphous film under non-sink conditions ilevbare2013a P4; indulkar2015 P13. (4) As the break point of flux against donor concentration [trasi2015a P6]. (5) By calculation from the crystalline solubility, the free-energy difference and the water activity of the amorphous drug ilevbare2013a E1. The papers of 2017–2022 add quantitative ¹H NMR of the dissolved drug beside the drug-rich phase [ueda2021 P3, P4], ultracentrifugation and the flux plateau in bile-salt media [lu2017b P3, P4], and an in-situ UV probe after a pH shift [sugihara2018 P6]. In the Bioperformance papers: (1) solvent-shift UV extinction, a methanol stock pumped into stirred buffer and the onset of scattering read at 350–455 nm [raina2014a P7; alhalaweh2016 P9; indulkar2016 P3; hate2018 P2; elkhabaz2017 P4]; (2) ultracentrifugation or centrifugation of a supersaturated solution and assay of the supernatant [mosqueragiraldo2018b P8; alhalaweh2016 P10]; (3) the flux plateau in a diffusion cell raina2014a R16; (4) estimate from the crystalline solubility with the Hoffman free-energy difference and a water-sorption correction raina2014a E3, E5; (5) the drug's own fluorescence, whose emission intensity rises, and whose peak may shift to shorter wavelengths, when the drug-rich phase forms, at the concentration found by ultracentrifugation in buffers and in pooled human intestinal fluid [elkhabaz2019b R3, C17]; (6) a solvent-shift or antisolvent addition into buffer with the polymer already dissolved, which gives the value in the presence of that polymer [ueda2020a R1; alkalla2026 R3, P7]; (7) for tacrolimus the value of about 50 µg/mL is taken from earlier work and used to define the dose-to-volume conditions of a dissolution test [hate2019 R7, C16; purohit2019 C15].

Values in the library
materialvalue or classconditionspaper ids
ritonavir37.2 µg/mL (onset); 39.2 predicted; 34.2 from filmwater, 37 °Cilevbare2013a R8, R13
ritonavir; lopinavir; paclitaxel hydrate26–28; 16–17.3; 27.2–30.1 µg/mL100 mM phosphate pH 6.8, 25 °C, four methods[trasi2015a R1]
telaprevir90–92 µg/mL (UV, NMR, fluorescence)pH 6.8, 37 °Cmosqueragiraldo2015 R13
clotrimazole; atazanavir7.7; 65.1 µg/mL (un-ionized)37 °C, 0.1 M bufferindulkar2015 R2
ibuprofen; sorafenib; griseofulvin125.74; 0.532; 355.96 µg/mLpH 2 or 6.8, 25 °C; griseofulvin with HPMCAS-HFalmeidaesousa2015 R5, R6
CRH1 free base12.49–13.73 µg/mLpH 4.6 citrate + 20 µg/mL HPMC, 25 °C[almeidaesousa2016 R4]
atazanavir70 µg/mL (onset); 68 predicted; with 3.6 mg/mL SDS about 1450 µg/mL total but 65–70 µg/mL freepH 6.8 buffer, 37 °C[indulkar2017a R5, R6, R11, R13]
telaprevir146–149 µg/mL by three methods; 311–360 with 12 mM STDC or SGDC50 mM pH 6.5 buffer, 37 °C[lu2017b R2]
pazopanib19.8 µg/mL at pH 6.5; 9.2 at pH 8.4; 60 at pH 50.1 M buffers with 5 µg/mL HPMC, 37 °C[sugihara2018 R1]
ibuprofen177.1 (buffer); 96.9 (HPMC); 90.7 (PVP-VA); 256.3 µg/mL (EUD-E)pH 2.0 (D2O), 300 µg/mL polymer, 37 °C[ueda2020b R1]
ibuprofen154 µg/mL (buffer) → 89–131 µg/mL with 1000 µg/mL of six polymersNMR, pH 2.0 (D2O), 37 °C[ueda2021 R5]
ibuprofen in bufferabout 127 µg/mL at 15 °C, about 284 at 60 °CNMR, pH 2.0 (D2O)[ueda2022 R9]
felodipine5.99 / 8.70 µg/mL predicted; onset 6.01 / 8.70pH 6.8 buffer, 25 / 37 °Craina2014a R16
nifedipine72.09 µg/mL predicted; onset 72.85buffer with 1 mg/mL HPMC, 37 °Craina2014a R16, X3
felodipine, FaSSIFonset 214.6 µg/mL; estimate 308.037 °C[raina2015c R1]
ritonavir / atazanavir / lopinavir45.2 / 132.1 / 27.2 µMpH 6.8, 37 °C[alhalaweh2016 R6]
clotrimazole7.5 µg/mL (ratio 15)pH 8, 37 °C[indulkar2016 R2, R3]
atazanavir78 µg/mL (94 with 1.8 mg/mL PVPVA)pH 6.8, 1 mg/mL HPMCAS[hate2018 R2, R3]
ezetimibe21.0 µg/mL buffer; 123 µg/mL FeSSIF37 °C[elkhabaz2017 R3]
telaprevir96–111 µg/mL with 1.05 mg/mL polymerpH 6.8, 37 °C[mosqueragiraldo2018b R13]
tacrolimusabout 50 µg/mL (cited), ratio about 35USP medium I[purohit2018 R4; trasi2016 C16]
enzalutamideonset about 42 µg/mL (crystalline 2.9); the printed estimate of about 39 is not reproduced50 mM phosphate pH 6.5, 37 °C[wilson2018 R1, R2, R3, C16]
atazanavir, clozapine, ketoconazole81.5 / 233.0 / 75.0 µg/mL without polymer; with AS-HF 35.5 / 95.0 / 56.0; with AS-LF 75.8 / 219.0 / 71.2buffer, 37 °C, 1000–2700 µg/mL polymer[ueda2020a R1, C13]
nifedipine123 µg/mL with AS-HF against 221 with AS-LFbuffer, 37 °C, 1000 µg/mL polymer[ueda2020a R1]
atazanavir, darunavir93.1 ± 1.5 (crystalline 1.6) and 363.8 ± 2.1 µg/mL (hydrate 265.6)pH 6.8, 37 °C, antisolvent[alkalla2026 R3]
atazanavir, posaconazole79–102 and 8.1–43.3 µg/mL in six media (crystalline 1.4–7.1 and 0.22–3.2)FaSSIF-type media and human intestinal fluid pool, 37 °C[elkhabaz2019b R1, R2, C15, C16]
pretomanidabout 166 µg/mL in buffer, 226 with SLS, 275 with TPGS, 308 in FaSSIF; crystalline 15–18, ratio 8–10pH 6.5, 37 °Cnguyen2023a R1, R2
tacrolimusabout 50 µg/mL, crystalline 2.5 µg/mL, ratio about 33used to fix the dose-to-volume conditions[hate2019 R7; purohit2019 C30]
Pitfalls and disagreements

The onset depends on the method by up to about 25% (ritonavir in water at 37 °C, 34–40 µg/mL) and on the medium (water against 100 mM buffer, twofold), so values are comparable only within one medium ilevbare2013a X14, X16, C40. Four of the eight 2013 predictions cannot be recomputed from the printed inputs ilevbare2013a X1. The claim that the onset equals the amorphous solubility is partly circular in the free-energy comparison almeidaesousa2015 X4. For ionizable drugs the value is for the un-ionized form indulkar2015 C47. For ibuprofen the same medium gave 177.1 µg/mL by extinction and 154 µg/mL by NMR, with crystalline solubilities of 29.8 and 43.1 µg/mL, in two papers of one group [ueda2020b R1, X1; ueda2021 R5, R2]. In SDS a prediction from the crystalline solubility overshoots about twofold [indulkar2017a C16]. For a drug that degrades on melting (pazopanib) the calculation cannot be made [sugihara2018 C17]. In the Bioperformance papers the onset depends on stirring, addition rate and polymer raina2014a X15. The estimate fails in FaSSIF and for high-melting telaprevir [raina2015c C18; mosqueragiraldo2015 C24]; the nifedipine prediction cannot be reproduced raina2014a X3. The atazanavir crystalline solubility is 5 µg/mL in one card and about 1.1 µg/mL in two others, so ratios differ [hate2018 R1; alhalaweh2016 R6; indulkar2017a E3]. A second drug or a polymer in the drug-rich phase lowers it (F6, F7). The value depends on the medium and on the polymer in it: the more hydrophobic HPMCAS grade AS-HF lowered it for four drugs by a quarter to well over half, and a co-dissolved crystalline drug lowered it too (F19, F20) [ueda2020a R1, C13; alkalla2026 C1, C39]. In simulated and human intestinal fluids the amorphous solubility rose much less than the crystalline solubility, so the ratio of the two fell from about 36 to about 5 for posaconazole (F18) [elkhabaz2019b R1, R2]. The printed enzalutamide estimate cannot be reproduced from its inputs [wilson2018 R3, X1]. The amorphous solubility of atazanavir at 37 °C is 75 (cited), 79, 81.5 and 93.1 µg/mL in four later papers, with crystalline values of 1.4, 1.14 and 1.6 µg/mL, so the ratio is 56, 71 or 58 depending on the paper [hate2022 M1; elkhabaz2019b R1, R2; ueda2020a R1; alkalla2026 R3] (synthesis: 79/1.4, 81.5/1.14, 93.1/1.6).

← Concepts and formulas

concept · as used in Crystallization tendency

antiplasticization

The rise of the Tg of a drug on mixing in a polymer of higher Tg, and with it a lower mobility, proposed as a reason for the stabilization of amorphous dispersions taylor1997 C4; kestur2011 M3. PVP, with a Tg of 177 °C, "has been suggested" to raise the Tg of the mixture and lower the mobility taylor1997 C4, M4. For indomethacin, inhibition by PVP at relatively low levels was already known to be too large to be explained by antiplasticization alone taylor1997 C5. In the growth studies of this chapter, antiplasticization is ruled out as the cause at 3–5 wt% polymer, since the dispersion Tg changes by 2.2 °C or less kestur2010a C32; kestur2011 C12. New aspect (Physical stability): water undoes it: after 75% RH every felodipine dispersion with up to 25% polymer had a Tg below the dry drug konno2008 C16.

Findings

Miscibility and drug-polymer interactions: F11, F1. Physical stability: F5, F6.

Related concepts

depends on and is measured by glass transition temperature; contrasts with drug-polymer interactions as an explanation of crystallization inhibition; related to supercooled liquid; contrasts with plasticization and plasticization by water.

Formulas

Gordon-Taylor equation.

How it is measured or obtained

By onset Tg of dispersions by DSC at 10 °C/min after erasing thermal history, against the drug alone and the pure polymer kestur2010a P8; kestur2011 P5. Growth rates may be normalized to Tm − T and T − Tg to separate driving force and mobility kestur2010a C31, E2. DSC Tg against polymer content, dry and after humid storage konno2008 P6; marsac2006a R17.

Values in the library
materialvalue or classconditionspaper ids
felodipine with 3% PVP, PVP/VA, HPMCAS, PVAcTg 44.1, 43.7, 42.7, 43.5 °C against 43.3 °Conset DSCkestur2010a R1
bifonazole with 5% polymersTg 16.4–17.4 °C against 15.2 °Conset DSCkestur2011 R6
nimesulide with 5% polymersTg 19.9–20.8 °C against 19.5 °Conset DSCkestur2011 R6
felodipine with PVP, HPMC, HPMCAS (other chapter)PVP raised Tg by 10 °C at 25 wt%; HPMC and HPMCAS no change up to 25 wt%onset DSCkonno2006 R6, C25
felodipine with PVP (other chapter)Tg largely unaffected (43.4–44.1 °C against 43.3 °C at 0.5–4.5 wt% PVP); antiplasticization not the cause, yet growth slowedPVP grades, 3–4.5 wt%; drykestur2010b R1, C34
felodipine–PVP 10 wt%Tg 40.5 °C, below pure felodipine 43.1 °Cdrymarsac2006a R8, C22
felodipine with ≤ 25% polymerTg ≈30–35 °C after 75% RH25 °Ckonno2008 R7, C16
celecoxib with 40% PVP K12 or PVPVA vs HPMCAS or HPMCdry Tg 84.4 and 81 °C vs 64 and 71 °C, although neat PVP K12 and PVPVA have lower Tg (97, 102 °C) than HPMCAS and HPMC (117, 140 °C)DSC, dry[xie2016b R17, R14, C27]

Evidence table: antiplasticization.

Pitfalls and disagreements

For nimesulide the inhibition order follows the polymer Tg, and the rank of dispersion Tg matches the inhibition rank for both drugs, within about ± 1 °C kestur2011 X8. At higher polymer contents the Tg does rise, and PVP is the strongest antiplasticizer of the three polymers in another study konno2006 C25. The authors conclude that Tg is not a good predictor of crystallization tendency in multicomponent amorphous systems kestur2010a C33. PVP raised felodipine Tg only from about 15% polymer, although the authors write "at all concentrations" konno2006 X15. The dry Tg gain did not carry over to humid storage: after 3 days at 94% RH four celecoxib dispersions had Tgs of 37.4–42.1 °C and very different stability [xie2016b R17, C46].

← Concepts and formulas

concept · as used in Crystallization tendency

atomic layer coating

Deposition of a thin metal-oxide film on the surface of powder particles by repeated, self-limiting vapour-phase reactions in a rotating reactor. For Al2O3, trimethylaluminum reacts with surface hydroxyl groups and water regenerates them, giving overall 2 Al(CH3)3 + 3 H2O → Al2O3 + 6 CH4 duong2022c E5, E6, E7. In the ASD process papers it is applied at 35 °C to amorphous solid dispersion (ASD) particles made by spray drying, by rotary evaporation or by extrusion and milling, with Al2O3 in all four coating papers and ZnO in one [duong2022c P2, P4; moseson2022a P1, P2, P3; benson2025 P3]. The growth per cycle was varied from slow (under 0.4 nm per cycle) to average and fast, and thickness is reported in nm [moseson2023 P3, P4; benson2025 P3]. The coating was introduced to stop surface crystallization of ASD powders; it also changes flow, water uptake and release.

Findings

F42, F43, F44, F45, F46, F47, F48, F49.

Related concepts

characterised by coating conformality and coating defect density; acts through surface molecular mobility and the moisture barrier effect; targets surface crystallization; sized with the specific surface area; changes powder flowability.

Formulas

coating thickness from oxide content and specific surface area.

How it is measured or obtained

Thickness: residual oxide after heating the coated powder in dry air (800–850 °C, thermogravimetry, TGA) converted with the BET area (see coating thickness from oxide content and specific surface area), and cross-sectional TEM of focused-ion-beam lamellae of about 100 nm moseson2022a P9, P10, R2; duong2022c P5, P10, R2. Effects: SEM, XRPD and DSC of powders stored open at 40 °C; dynamic vapour sorption; shear-cell and tapped-density flow tests; release in non-sink tests; SEM/EDX of particles incubated in medium moseson2022a P11, P12, P14; duong2022c P8, P9, P15, P16; moseson2023 P10, P13, P15.

Values in the library
materialvalue or classconditionspaper ids
ezetimibe–HPMCAS 50 and 70% DL, spray dried, thin and thick Al2O3about 12–40 nm by TEM (six places in one cut); oxide residue 1.7–9.2%rotating-tube reactor, 35 °Cduong2022c R2, R12
erlotinib–HPMCAS, erlotinib–PVPVA, naproxen–HPMCAS, lumefantrine–PVPVA7.4–24 nm by TGA and BET; 4.4–37.7 nm by TEM; Al2O3 and one ZnOrotary reactor, 35 °Cmoseson2022a R2
posaconazole–HPMCAS 25 and 50% DL, twelve samples11–39 nm (oxide 0.40–3.68%); one slow-deposited 26 nmslow, average, fast growthmoseson2023 R1
ritonavir–PVPVA 10 and 20% DL, nine coated samples9.3–28.3 nm; oxide 0.92–5.30%; BET 0.22–0.76 m²/g[benson2025 R1]
stored powders, uncoated → coatedcrystals by SEM within 1–3 weeks → 3–24 weeks, or none within 48 weeks to 2 yearsopen dishes, 40 °C, 31–75% RHmoseson2022a R11; duong2022c R16, R17
tablets of coated ritonavir–PVPVA particlesabout 35–75% of the dose at 60 min against about 95% uncoatedpH 6.8, 37 °C[benson2025 R10]
Pitfalls and disagreements

The thickness label is a mass-balance estimate and differs from TEM by up to a factor of two inside one paper moseson2022a X10. The number of cycles is not printed in the coating papers [moseson2023 P3; moseson2022a P3; duong2022c X2; benson2025 M11]. The coating delays crystallization and does not stop it, and it can slow or block release depending on defects, not thickness (see coating defect density) [moseson2022a R11; moseson2023 C53; benson2025 C21]. Evidence is for Al2O3 on six drug–polymer pairs; other oxides, polymers and dosage forms were not tested duong2022c C60; moseson2022a X20.

← Concepts and formulas

concept · as used in Crystallization tendency

Barandiaran-Colmenero method

A DSC method from inorganic glass science to estimate the critical cooling rate: the natural logarithm of the cooling rate is regressed on 1/\Delta T^2, where ΔT is the undercooling at the crystallization peak, and Rcrit is taken as \exp(A), the extrapolated rate at which no crystallization would be seen baird2010 E1, E2; baird2013 E1, E3. The Cabral variant subtracts B/T_m^2 from the intercept baird2010 E3.

Findings

F11.

Related concepts

estimates critical cooling rate; uses degree of supercooling; related to glass-forming ability.

Formulas

Barandiaran-Colmenero critical cooling rate, Cabral critical cooling rate.

How it is measured or obtained

Crystallization peak temperatures at several cooling rates, 5–20 °C/min in a conventional DSC baird2010 P2, P9 or 5 to about 800–1100 °C/min with rapid heat–cool DSC baird2013 P3.

Values in the library
materialvalue or classconditionspaper ids
carbamazepineRcrit 118 K/min, r² 0.995–20 °C/minbaird2010 R13.j
griseofulvin588 K/min, r² 0.975–20 °C/minbaird2010 R13.m
4-biphenylmethanol3.28 × 10⁴ K/min, r² 0.745–20 °C/minbaird2010 R13.b
12 of 19 class I moleculesr^2 < 0.55, no estimate5–20 °C/minbaird2010 R13
phenacetin; benzocaine2.7 × 10³; 6.2 × 10³ K/min5 to ≈1000 °C/minbaird2013 R7.h, R7.d
benzamide, lidocaine, indoprofenr^2 < 0.555 to ≈1000 °C/minbaird2013 R7
Pitfalls and disagreements

The method needs undercooling to rise steadily with cooling rate, which often failed because of scattered crystallization temperatures baird2010 C18; baird2013 C17. Estimates from 5–20 °C/min contradicted liquid-N2 quench results baird2010 C21; accurate predictions are said to need rates at least two orders of magnitude below Rcrit baird2010 R31.

← Concepts and formulas

concept · as used in Crystallization tendency

Burger-Ramberger rules

Thermodynamic rules for polymorph pairs; the heat-of-fusion rule used in the library says that if the higher-melting form also has the larger heat of fusion, the pair is monotropic lu2016 E3; trasi2014b E2. Neither paper prints the rule itself; both apply it in words.

Findings

F7.

Related concepts

tests monotropy; part of polymorphism; uses enthalpy of fusion.

Formulas

Burger-Ramberger heat-of-fusion rule.

How it is measured or obtained

From the DSC melting points and enthalpies of fusion of both forms lu2016 R20; trasi2014b M1, R6.

Values in the library
materialvalue or classconditionspaper ids
vemurafenib α against β271 °C, 53.8 kJ/mol against 248 °C, 33.3 kJ/mol: monotropicDSClu2016 R20, C50
nilutamide stable form against form Iaround 153 °C, ~98 J/g against 141 °C, around 66 J/g: monotropicDSCtrasi2014b M1, R6, C11
Pitfalls and disagreements

The vemurafenib β enthalpy may be a lower bound because recrystallization overlaps its melting, and the α value differs between parts of the paper lu2016 X12, X1; the conclusion would change only if β's enthalpy reached α's lu2016 X12.

← Concepts and formulas

concept · as used in Crystallization tendency

classical nucleation theory

The theory that writes the nucleation rate as a pre-exponential factor times \exp\left[-\frac{W^* + \Delta G_\eta}{kT}\right], with a thermodynamic barrier W^* = \frac{16\pi\sigma^3}{3\,\Delta G_v^2} for a spherical nucleus and an activation energy for viscous flow baird2012 E1, E2; vaneerdenbrugh2010a E3. Its main factors are driving force, interfacial energy and mobility, and the library adds molecular recognition trasi2012a C48; konno2006 C33. In this chapter it is used qualitatively; solution papers fit it to induction times. New aspect (Crystallization from solution): For nucleation from supersaturated aqueous solution the papers use the form J = A\exp(-\Delta G_{crit}/kT) with a barrier that falls steeply with supersaturation, \Delta G^{*} \propto 1/(\ln S)^{2} (a pre-exponential factor A that depends on molecular mobility); the induction time is then inversely proportional to J if growth is fast hsieh2012 E7, E8; ilevbare2013c E3, E4. Two further elements are added: the polymer's effect through the interfacial tension or the cluster reorganization, and the two-step picture in which a disordered solute cluster forms first and then reorganizes, the reorganization being taken as rate-limiting ("we assume") ilevbare2013c C5, C15. The acetaminophen study prints J = A\exp[-16\pi\gamma^3 v^2/(3k^3T^3(\ln S)^2)] to say that temperature, supersaturation and interfacial tension set J, and fits a Poisson model to induction-time distributions for J trasi2015 E6, C20, E5. From 2018 the theory is also used to read medium effects: a lower interfacial energy for lecithin, SDS hemimicelles and phospholipid digests, a lower attachment rate in the pre-exponential factor for human fluid, and a fit of \ln(J/S) = \ln A - B/\ln^{2}S to posaconazole induction times in buffer and four media [indulkar2018 C40; indulkar2019b C33; enright2020 C26; elkhabaz2021 C22; duong2022b E7]. New aspect (Physical stability): the physical-stability papers use the theory to sort possible polymer effects into the thermodynamic barrier (driving force, interfacial energy) and the kinetic barrier, and conclude that a few percent of polymer acts on the kinetic barrier konno2006 C34, C36, C40; marsac2006a C44, C48. New aspect (Supersaturation from ASD dissolution): used qualitatively for dissolving dispersions: nucleation rises steeply with supersaturation, so amorphous–crystalline systems with a high theoretical solubility ratio are expected to nucleate readily [konno2008a C20]; at equal supersaturation in two coexisting phases nucleation is favoured in the drug-rich droplets, where the drug concentration is higher and the crystal–liquid interfacial tension lower [ilevbare2013d C40, E1]. A celecoxib paper of 2015 prints the classical rate with (\ln S)^2 and argues from it that the polymer differences come neither from supersaturation, taken as similar in all polymer solutions, nor from interfacial tension, which adsorbed polymer would lower and so speed nucleation; the theory therefore "does not readily allow for obvious mechanistic insights", and two-step nucleation through dense liquid clusters is offered instead (cited) xie2015 E3, C43, C44, C45. The longer celecoxib induction time at 11 than at 22 µg/mL is read as in agreement with the theory [xie2016a C13].

Findings

Crystallization tendency: F3, F6. Crystallization from solution: F1, F5, F13, F14, F27, F34, F39, F42. Physical stability: F2, F4, F17. Supersaturation from ASD dissolution: F4, F5, F9, F13, F22.

Related concepts

models nucleation; depends on thermodynamic driving force for crystallization and melt viscosity; contrasts with heterogeneous nucleation; related to prenucleation clusters; governs nucleation rate and nucleation induction time; depends on interfacial tension; related to crystal-solution interfacial free energy; includes kinetic barrier to nucleation and interfacial free energy; depends on supersaturation.

Formulas

classical nucleation barrier, critical cooling rate to avoid nucleation, classical nucleation theory, induction time decomposition.

How it is measured or obtained

Not fitted for melts in this subcategory: the classes are read through it, a more negative ΔGv giving a lower barrier baird2010 C61, interfacial energy differences being assumed small vaneerdenbrugh2010a C39, and a steeper viscosity rise lowering the rate baird2012 C32. In solution, the barrier and interfacial energy are fitted from induction times against supersaturation duong2022b E7, E8. Most of the solution-crystallization papers do not fit it; they use it qualitatively to explain why induction times fall with rising supersaturation and why a polymer gains more at a lower one (celecoxib 27.5 min at S 7.3 against 2.4 min at S 14.7) ilevbare2013c R15, R9.a, C45. The posaconazole paper converted induction times into nucleation rates (conversion not stated) and fitted the high-supersaturation points of each medium with ν = 920 ų and T = 310 K duong2022b P15, X10. In the 2006 felodipine and nifedipine film papers it is used as a framework only; no parameter was fitted konno2006 E8; marsac2006a E9. In the ASD dissolution papers no rate was computed; the expression is printed in two papers and stated in words in a third [konno2008a E3; alonzo2010 E4; ilevbare2013d E1].

Values in the library
materialvalue or classconditionspaper ids
posaconazoleinterfacial energy 3.52 mJ/m²; ΔG* ≈2.73 × 10⁻²¹ JPBS, 310 K, fit to induction timesduong2022b E7, E8
classes I against IIIlower barrier inferred for class I from more negative ΔGvmelt classes, γ assumed similarbaird2010 C61
felodipine filmsrate described by driving force, kinetic barrier and molecular recognition; nucleation most likely heterogeneous0% RH filmskonno2006 C33
celecoxibinduction time 27.5 ± 0.7 min at S 7.3; about 2.4 min at S 14.737 °Cilevbare2013c R15, R9.a
danazol with HPMC-AS516, 365, 277 min at 10, 15, 20 µg/mL25 °C[jackson2014 R17]
type 1 weak basesprecipitate only after a supersaturation of 1.4–26.7 times the titration solubilitypH shift, 37 °Chsieh2012 R5, C3
celecoxib dispersions (ASD dissolution chapter)nucleation-rate expression with \ln S and interfacial tension \gamma used to read induction times37 °Cxie2015 E3, C42
acetaminophen, S 1.5, 20 °CJ 1465 (none), 3820 (PAA), 81.86 (HPMCAS), 40.68 (HPMC) # m⁻³ s⁻¹ for V = 1.2 mL10 µg/mL polymertrasi2015 R3, X1
posaconazole, PBS, FaSSIF V1, FaSSIF V2, FeSSIF V1γ 3.52, 2.61, 2.73, 3.24 mJ/m²; A 2.44, 5.14, 4.98, 9.87; R² 0.84–0.9737 °C, high-S points (3–9 per medium)duong2022b R15, R16, X11
posaconazole, PBS, S 13 → 33.6ΔG* about 5.11 → 2.72 × 10⁻²¹ J; r* about 5.89 → 4.29 × 10⁻¹⁰ m310 Kduong2022b R17
seven other drugs (cited)γ 0.60–2.80 mJ/m²literatureduong2022b R18
felodipine with 3–25% polymerthermodynamic route "probably cannot account" for the drop; kinetic barrier speculatedargumentkonno2006 C35, C40
nifedipine vs felodipinelarger enthalpic driving force and lower barrier below Tg for the faster nucleatorTable II, Table Imarsac2006a C49
flutamideprenuclei redissolve unless they exceed a critical radius, the volume free energy against the interfacial termframeworktrasi2012b C46
felodipine (argument)dissolution of the amorphous drug "will be accompanied by rapid crystallization"37 °C[konno2008a C21]
felodipine 90:10 dispersion (argument)almost all drug enters a highly supersaturated state that immediately crystallizes ("We speculate")37 °C[alonzo2011 C29]
celecoxib, no polymer, 22 against 11 µg/mLonset within 5 min against about 20 minpH 6.8, 37 °C[xie2016a R2]
celecoxib 22 µg/mL, 5 µg/mL PVP-K12 / PVP-K29/32 / HPMCAS or HPMConset ≈1 h / ≈2 h / more than 8 hpH 6.8, 37 °Cxie2015 R1
paclitaxel, critical radius of a two-dimensional nucleus5.4–7.0 nm at S = 88 with three stand-in interfacial tensions; ≈6.8–8.8 nm at the residual supersaturation after 16 hpH 6.8, 37 °C[que2018 R15, R17]
Pitfalls and disagreements

Pharmaceutical films are not completely described by homogeneous nucleation theory, which ignores stresses and heterogeneous nucleation marsac2006a C32. It does not readily explain polymer effects on nucleation or encompass molecular recognition trasi2012a C49; xie2015 C44. In the melt papers neither γ nor nucleation rates were measured, so the barrier argument is an inference vaneerdenbrugh2010a C39; baird2010 C61. The proportionality of induction time to 1/J assumes nucleation is much slower than growth to detectable size, which was not tested ilevbare2013c X14. The two-step model is assumed, not shown, for these systems ilevbare2013c C5. The critical supersaturation at which precipitation is seen depends on the compound and the conditions and is somewhat variable hsieh2012 C3. The printed sample volume (0.0012 m³) and the sign of the exponent in Eq. 5 are misprints trasi2015 X1, X2. The human-fluid paper prints the rate equation without the minus sign in the exponent [elkhabaz2021 X1]. In the posaconazole fits γ and A both differ between media and the critical free energy in the FaSSIF media is less than half the buffer value at equal S, so the assignment of the medium effect to A alone is not secure duong2022b X12. In the felodipine films the homogeneous theory is applied to nucleation that is "most likely heterogeneous" konno2006 C33; marsac2006a C32. The reduced-temperature form printed for acetaminophen has typesetting errors (r³ for γ³, a positive kinetic term) trasi2012a X1. The 2008 and 2010 dissolution papers print ln S without the square of the classical form [konno2008a X8; alonzo2010 X3]. The celecoxib supersaturation argument uses crystalline solubilities measured with 22 µg/mL polymer, not the 5 µg/mL of the induction tests, and these differ by 26–43% from the polymer-free value xie2015 X9; the text refers to the wrong equation for interfacial tension xie2015 X1. The paclitaxel values use interfacial tensions of other compounds, and with the printed units the equation gives a dimensionless number [que2018 C37, X7].

← Concepts and formulas

concept · as used in Crystallization tendency

cold crystallization

Crystallization on heating a glass into its supercooled liquid (devitrification), as opposed to melt crystallization on cooling lu2016 C2. In the DSC classification it separates class II (crystallizes on reheating) from class III baird2010 R1.

Findings

F7, F15.

Related concepts

process in supercooled liquid; contrasts with crystallization from the supercooled melt; measured by critical heating rate; related to glass stability and polymorphism.

Formulas

none.

How it is measured or obtained

As an exotherm on DSC reheating of a quenched glass, or by isothermal holds above Tg on a hot stage lu2016 P2, P3; baird2010 P1; the slowest heating that avoids it is the critical heating rate baird2013 R9.

Values in the library
materialvalue or classconditionspaper ids
vemurafenib glassexotherms peaking at 157 and 217 °C, then α melting at 271.5 °CDSC at 10 °C/minlu2016 R3
11 class II moleculescrystallize on reheating at 10 °C/minDSCbaird2010 R1
nifedipine glasscrystallized to a metastable form at 87 °C; felodipine did notheating at 2 °C/minmarsac2006a R5
delamanid glassdevitrified just above Tgliquid-N2 quenchedduong2022a R3
propranolol saltsrecrystallization at 46.5–105.4 °C for four salts; none for nicardipine saltsDSCtowler2008 R4, C10
rotary-evaporated drug–polymer powdersno exotherm on heatingDSCvaneerdenbrugh2010b R10
Pitfalls and disagreements

The polymorph formed on heating is often metastable and may convert later lu2016 C54. The rate at which the glass was made changes cold crystallization on reheating baird2013 C28.

← Concepts and formulas

concept · as used in Crystallization tendency

conformational flexibility

The ability of a molecule to take different shapes by rotation about single bonds, counted in the library as the number of rotatable bonds baird2010 P8. More flexible molecules exist in the melt as many similar conformers, lowering the probability of the orientation needed to nucleate or grow, which is argued to raise glass-forming ability baird2010 C59; vaneerdenbrugh2010a C37. New aspect (Crystallization from solution): in solution crystallization the argument is turned around from the melt picture, as molecules with several low-energy forms (conformers or tautomers) are said to crystallize more slowly because only the right form joins a nucleus, which lowers the effective supersaturation of that form hsieh2014 C31. Ilevbare and co-workers attribute the slow nucleation of ritonavir, the largest molecule (720.9 g/mol), "most likely" to its higher molecular weight and greater conformational flexibility ilevbare2013c C25, M1. Among six dihydropyridines, rotatable bonds (6–11), molecular weight and polar surface area did not track the order of crystallization from the precipitate, and only cilnidipine stands out as larger and more flexible raina2015b C31, M1–M6.

Findings

Crystallization tendency: F7, F13. Crystallization from solution: F3, F16, F21.

Related concepts

related to entropy of fusion and polymorphism; contributes to crystallization tendency; related to polymer chain stiffness.

Formulas

none.

How it is measured or obtained

Rotatable-bond counts from software or databases baird2010 P8; baird2012 P7; single-molecule conformer energy scans for one drug trasi2014b P10. In the solution papers not measured; rotatable-bond and tautomer counts are quoted: dipyridamole 12 rotatable bonds, warfarin 4; tautomers 6 (glyburide) and 9 (warfarin) hsieh2014 X12, R19.

Values in the library
materialvalue or classconditionspaper ids
class I against class IIIrotatable bonds ≈−0.54 against ≈+0.66 SD51 moleculesbaird2010 R22.a
solvent classes I against III≈−0.42 against ≈+0.57 SD40 drugsvaneerdenbrugh2010a R20.b
ritonavir18 rotatable bonds, class IIImeltbaird2012 M34
nilutamideone rotatable bond; crystal conformer at 54.4° against a computed minimum near 30°DFT, crystal structuretrasi2014b M1, R19, R24
ritonavir in solution, against celecoxib and efavirenzslow nucleation attributed to higher MW and flexibility; induction time about 120 against 2.4 and 5.1 minaqueous buffer, 37 °Cilevbare2013c R8, C25
glyburide, warfarinseveral tautomers may explain slow crystallization; amorphous precipitates for at least 6 h with most polymerspH shift, 1 mg/mL polymerhsieh2014 R4, R19, C31, C32
papaverine, dipyridamolereduced conformational freedom proposed; crystalline with every polymer; papaverine adopts its crystal conformation in solutionpH shifthsieh2014 R4, R19, C33
six dihydropyridinesno correlation of rotatable bonds with the first crystalline time (0.25–12 h)10 mg/mL solvent shiftraina2015b R6, C31
Pitfalls and disagreements

Conformer populations were never measured; the argument is inferred from class averages baird2010 C59. Nilutamide, with a single rotatable bond, is nonetheless a good glass former trasi2014b C36. In one solution study the rotatable-bond counts did not follow the conformational-freedom ranking used for dipyridamole and papaverine, and the argument rests on cited data hsieh2014 X12. Neither the ritonavir explanation nor the tautomer explanation was tested ilevbare2013c C25; hsieh2014 C31. Prediction from structure is called extremely difficult by the authors of the dihydropyridine study raina2015b C6.

← Concepts and formulas

concept · as used in Crystallization tendency

congruent release

Drug and polymer leave a dispersion at the same normalised rate, so release is governed by the polymer; typical of low drug loading, and the condition for fast, complete release of a copovidone dispersion [li2020 C63]. Above a drug-specific loading release becomes incongruent and falls to roughly that of the neat amorphous drug (see limit of congruency). In the bicalutamide dispersion, congruent release at 5 and 10% DL went with escape from matrix crystallization; the authors conclude that it "may be a strategy to avoid" it, yet a congruently releasing dispersion with 3% SDS still lost one of three replicates moseson2022b C36, C48, C51. In the Peptide permeability papers: The criterion extends from two components to three, and from a small molecule to a peptide with its permeation enhancer: in an octreotide–SNAC–copovidone dispersion all three components released at one normalized rate, which the authors call synchronous release [bapat2025b C3, C15]. It also survives a component that is not amorphous: both enhancers stayed crystalline in the dispersions and were still carried out with the polymer, which the authors attribute "likely" to the gel-forming properties of copovidone [bapat2025b R17, C44]. A second usage appears in the same subcategory under the name corelease, judged not by rates but by the times at which each component reaches its maximum release from a tablet, which were about 30 min for insulin and about 10–20 min for the enhancer [fagan2025 R20, C44, C46]; see peptide-enhancer co-release. New aspect (Release mechanism and LoC): in the release-mechanism papers of 2017–2020 congruent release is read from drug and polymer assayed separately from a compact of constant area, as equal rates normalized to each component's weight fraction (no significant difference by t-test) or as overlapping percent-release curves [saboo2019 R8, E4; indulkar2019a R3]. Congruent dispersions keep releasing into a medium presaturated with drug, which the authors take as proof of polymer control [indulkar2019a R12, C43; indulkar2017b C35]. The 2017 ritonavir study ties the colloid to release that is "congruent or close to congruent" [purohit2017b C52], and the 2019 dihydropyridine study calls congruent release the "sweet spot" of dispersion dissolution because only it gave drug-rich droplets ("a clear link") [saboo2019 C30]. Congruent release ends at the limit of congruency, which lay between 5 and 30% drug in PVPVA, and with HPMCAS or Eudragit S 100 it lasted to 50% felodipine [que2019 C10; indulkar2019a C2; saboo2020a R15]. The papers of 2020–2022 add that congruent release can end after minutes when crystals form on the dissolving surface (indomethacin methyl ester–PVPVA congruent at 5 min up to 25%, over 30 min only at 5%) saboo2020 R3, R8, C17, that at the limit the polymer controls release whatever the compound [que2021 C39], and that congruent release at a limit raised by a surfactant is slower than at lower loadings (≈1.8 against ≈2.95–3.2 mg/min/cm² for felodipine with 5% TPGS) [saboo2021 X3]. For ritonavir at 30%, one loading above its limit, every surfactant built into the dispersion removed the lag phase and gave congruent release for at least 20 min, but only Span 85 and TPGS kept it congruent to about 87–95% at 90 min indulkar2022 C31, X14. For two phenolphthalein analogues in PVPVA the morphology of the two phases after separation "appears" to differentiate congruent from incongruent release; during congruent release the drug is carried into the diffusion boundary layer by swelling and release of the polymer chains [deac2023a C46, C47]. Below the limit the surface drug/polymer ratio of PHPH (5%) and MePHPH (30%) compacts stayed constant during dissolution, as it must if both components release in the dispersion ratio [deac2023b C24, C58]. The papers of 2023–2025 extend the reading to HPMCAS, where indomethacin, its ester and four weak bases released drug and polymer at matching normalized rates, so that release "appear[s] co-dependent" [bapat2024 C10, C23], and to PVP, where congruency was counted as more than 80% of both components and congruent dispersions released more slowly with higher-molecular-weight grades [deac2024a C36, C19]. In the PVP paper droplets in the gel are held until the polymer is dilute enough to leave and then leave with it, so congruent release is limited by water penetration and the release of chains [deac2024a C34].

Findings

ASD process: F21, F24, F38, F40, F48. Peptide permeability: F4, F5, F6. Release mechanism and LoC: F1, F2, F3, F4, F6, F11, F12, F13, F15, F17, F19, F21, F23, F24, F25, F26, F27, F29, F31, F32, F33.

Related concepts

ends at the limit of congruency; the opposite is incongruent release; belongs to polymer-controlled release; the three-component case for a peptide plus an enhancer is peptide-enhancer co-release; still bounded by the limit of congruency and opposed by incongruent release; the accelerating effect of a co-formulated enhancer connects it to water sorption of the excipient blend. Read by the intrinsic dissolution rate of each component; contradicts the Higuchi two-component dissolution model at low loading; the condition for drug-rich nanodroplets to form; ended early by surface crystallization; coupled to the polymer's own dissolution rate, which a cationic drug can lower through drug-polymer ionic interactions.

Formulas

The same surface-normalized rate is used, printed here as Eq. 1 with the medium volume in the numerator; see surface-normalized release rate [bapat2025b E1]. Also congruent release relation and saturated-medium condition.

How it is measured or obtained

Constant-area dissolution of compacts with drug and polymer both assayed, expressed as normalised release rates or as fractions released; in the ASD process papers the surface-normalised dissolution rate of ritonavir–PVPVA was used and the congruent boundary of 25% DL was cited from earlier work [li2020 P9, E13, R23]. In the Peptide permeability papers: As before by constant-area dissolution with every component assayed and converted to a normalized rate, here at 0.5 cm² in 100 mL of 50 mM pH 6.8 phosphate buffer at 37 °C, with reversed-phase HPLC for peptide and SNAC and aqueous size-exclusion chromatography for copovidone [bapat2025b P4, P2, E1]. The new practical limit is analytical: sodium decanoate could not be quantified at all, so in every decanoate dispersion congruence rests on the peptide and the polymer and the enhancer's co-release is inferred ("presumably") [bapat2025b R6, C19, X1]. In the release-mechanism papers: Wood's apparatus or a die with one face of about 0.5 cm², drug by HPLC or UV, polymer by size-exclusion chromatography, light-scattering detection or colorimetry, and the rate A = kV/(SN) for each component [indulkar2019a P2, P3, P4; saboo2019 P9, P10; saboo2020a E3, P7; bhardwaj2018 P6, P7, E4]. Later variants: 5-min rates sampled every minute saboo2020 P5; the last loading with simultaneous release and at least 80% of the drug in 90 min [que2021 P9]; t-tests of normalized rates [yang2021 P17]; drug release alone hiew2022b X6.

Values in the library
materialvalue or classconditionspaper ids
ritonavir–PVPVAcongruent up to 25% DL; at 40–50% DL drug release like neat amorphous drugcited[li2020 R23, C63]
ritonavir–PVPVA spray-dried, 25% DLrate about 3.5 mg·min⁻¹·cm⁻¹ from methanol, falling to about 0.90 with 60:40 water:methanolpH 6.5, 100 rpm[li2020 R10]
ritonavir–PVPVA, 50% and 75% DLabout 0.003–0.006 mg·min⁻¹·cm⁻¹, near neat amorphous drug (about 0.0026)same[li2020 R10]
ritonavir–PVPVA (outside)congruent up to 25% DL; polymer leaves first above ittablets[indulkar2019a C2]
felodipine ASDs (outside)PVP and HPMC congruent to 10% DL, PVPVA to 15%; HPMCAS and EUDS congruent to 50% DL37 °C[saboo2020a C12, C14]
BCL–PVPVA 5 and 10% DL, solvent evaporateddrug about 91 and 96% and polymer about 90 and 97% of the dose at 10 min; ends about 91 and 96%dose of at most 30 µg/mL, pH 6.8moseson2022b R6, R7
BCL–PVPVA 30% DL with 1.5% SDScongruent; more than 80% in 30 min, about 90–94% at 4 hsamemoseson2022b R9, R13
RTV–PVPVA 10 and 20% DL powders (below the limit of 25%)seven of eight coated powders about 94–97% at 60 minpH 6.8[benson2025 R4]
SNAC:copovidone 50:50congruent; ≈7.9 and ≈8.7 mg min⁻¹ cm⁻², about 4.4 and 2.7 times the neat rates; complete 25–30 minpH 6.8, 0.5 cm² compact, 37 °C[bapat2025b R2.d–e, R3, X18]
octreotide:SNAC:copovidone, 1.6% peptidecongruent in three components; ≈8.1 / ≈8.4 / ≈8.9 mg min⁻¹ cm⁻²same[bapat2025b R2.f–h, R5]
octreotide:sodium decanoate:copovidone, 1.6% peptide≈23 and ≈25 mg min⁻¹ cm⁻² (peptide, polymer); complete in about 10 min; enhancer not assayedsame[bapat2025b R2.j–k, R9, X1]
SNAC:copovidone ratio seriescongruent within 30 min from 20:80 upward; 10:90 slower, complete within 40 minsame (SI, text only)[bapat2025b R4, C14]
sodium decanoate:copovidone ratio seriespolymer complete within about 10 min from 20:80 upward, by 25 min at 10:90same (SI, text only)[bapat2025b R8, C18]
cyclosporine:copovidone ± enhancer, 1.6% peptidepeptide follows polymer: about 50 min (binary), about 10 min (decanoate), ≈95% at 60 min (SNAC)same[bapat2025b R10, C21]
cyclosporine ternaries, 10–20% peptidenot congruent: polymer complete while peptide reaches ≈54.5% and ≈16% with decanoatesame[bapat2025b R12.i, R13.i, C23]
nifedipine 10 and 20% with HPMC-E5 or PVPVAcomplete release in ~20 and < 5 min, also into water saturated with drug5 °C, D2O[indulkar2017b R8, R9, C35]
ritonavir 10:90 films, PVP, PVPVA, HPMCASdrug and polymer dissolved at the same rate; the polymer almost completely in < 10 min25 °C, pH 6.8[purohit2017b C52, R15, R16, R17]
ritonavir 50:50 HPMCAS filminitially congruent (≈32–34 µg/mL drug within 5 min)same[purohit2017b R17, C38]
ITZ–PVPVA 10% spray-driednormalized drug ≈2000 against polymer ≈2265 µg/min/cm²0.1 N HCl, 37 °C[bhardwaj2018 R11]
ritonavir–PVPVA 10, 20, 25% tablets≈97–100% of both by 60 minpH 6.8, 37 °C[indulkar2019a R3, R11]
ledipasvir–PVPVA 2 and 5%about 100% of both by 60 minpH 6.8, 37 °C[que2019 R3, R4]
ledipasvir–PVPVA with 5 parts SDS, 7.5–30%congruent to 30%; fails at 40%same[que2019 R7, R8, R9]
nilvadipine 5–10%, cilnidipine 5–15% in PVPVAdrug 2.13–2.53 = polymer 2.35–2.76 mg/min/cm² (ns)pH 6.8, 37 °C[saboo2019 R5, R6, R8]
felodipine with HPMCAS (10–50%) and EUDS (10–50%)drug = polymer at every loading; HPMCAS ≈0.13–0.93 mg/min/cm²pH 6.8; EUDS pH 7.4[saboo2020a R14]
nilvadipine, felodipine, cilnidipine 40% in HPMCAS0.42/0.42, 0.34/0.32, 0.26/0.24 mg/min/cm²same[saboo2020a R16]
indomethacin; its methyl ester in PVPVAIND 5, 10%: ≈3.44/3.47, 3.14/3.19; INDester 5–25%: ≈3.3–4.2 mg/min/cm² (drug/polymer, 5 min)pH 2.1 / 6.8, 37 °Csaboo2020 R3, M5, M6
indomethacin methyl ester–PVPVA 10–25%congruent at first, drug levels off after 7.5–20 min30 min, 37 °Csaboo2020 R8
four phenolphthalein analogues at their limitsdrug ≈2.9–3.7, polymer ≈2.8–3.4 mg/min/cm² (p = 0.022–0.67)pH 6.8 (TIPP 2.5)[que2021 R21, P9]
felodipine–PVPVA with 2, 5, 10% TPGScongruent to 25, 35, 45%pH 6.8, 37 °C[saboo2021 R12, P6]
loratadine–PVPVA; –HPMCASto 30% and 20% at 37 °C; 2% and 5% at 5 °Ccompacts, t-test[yang2021 R22]
clopidogrel–PVPVAclose to congruent to 40%; 50% with SDScompacts[correasoto2022a C46, C55]
atazanavir–PVPVA5%; 10% with SDS or CTAB; 15% with Span 80compacts[correasoto2022b C12]
ritonavir–PVPVA 30:70 with 5 parts Span 85, TPGS; SDS; Span 20; Tween 80congruent for at least 20 min with every surfactant; ≈87–89% (Span 85, TPGS), ≈78% (SDS), ≈41%, ≈25% of the drug at 90 minpH 6.8, 37 °Cindulkar2022 C31, R8, R9, R5, R7, R6
miconazole 20% on HP-50; on HP-50-Na; on PTEAincongruent and slow, drug ≈8.6% at 60 min; drug ≈89.5 and 83.7% at 60 min with the polymer aheadcompacts, 50 mM pH 6.8 phosphate, 37 °C[qi2022 R14, C22]
ritonavir–PVPVA 20%; 30%; 40–50%congruent and complete; drug ≈35% with the polymer ahead; drug ≤3% at 90 mincompacts, pH 6.8, 37 °C[yang2022a R2, C12, C13]
indomethacin–HPMCAS-MF 10%; its methyl ester 10%drug and polymer at the same normalized rate within error, ≈0.12 → 0.82 mg/min/cm² over 3–51 mM/ΔpH; rates comparablepH 6.8 phosphate and FaSSIF[bapat2023 R9, R10, R13, C16]
atazanavir–PVPVA 20% with TEAC, TBC, TBAC; at the limit, 25%drug ≈4.6–5.1 mg/min/cm²; ≈2.0–3.1 against neat PVPVA ≈2.7compacts, pH 6.8, 37 °C[correasoto2023 R6, R7, C13]
MePHPH–PVPVA 20, 40, 50, 70%; PHPH–PVPVA 5, 20%drug ≈96 / 61 / 15 / 0%; ≈75 / 0% at 30 mincompacts, pH 6.8, 37 °C[deac2023a R1, R2]
PHPH–PVPVA 5%; MePHPH–PVPVA 30% compactssurface drug content constant over 60 minATR-FTIR of dried surfaces[deac2023b R6, R7]
indomethacin, indomethacin methyl ester–HPMCAS-MF 5–60%drug ≈0.85 → 0.50 and 0.66 → 0.39 mg/min/cm² with the polymer alongsidecompacts, pH 6.8 phosphate, 37 °C[bapat2024 R2, R3, C10]
loratadine 25% in HPMCAS LF, MF, HFdrug/polymer ≈0.25/0.27, 0.10/0.12, 0.01/0.02compacts, pH 6.8[bapat2025a R2, C13]
clotrimazole–PVP K25 20%, K29/32 30%, K60 35%, K90 38%congruent (>80% of both components)compacts, pH 8.0, 90–120 min[deac2024a R1, R2]
ritonavir–PVPVA 30% + Span 20congruent for 20 min (≈48% drug, ≈50% polymer), then ≈92–97% drugcompacts, pH 6.8yang2023 R2, R3, C13
Pitfalls and disagreements

No numerical criterion is given for "congruent" in the ritonavir study [indulkar2019a X7]. Phase separation can raise or lower release depending on loading regime and on where the drug-rich phase ends up [li2020 C64]. The limit of the bicalutamide pair was not located and lies between 10 and 20% DL moseson2022b X15. A congruent SDS dispersion failed after milling moseson2022b C51. Peptide permeability: The loading boundary for a peptide is far lower than the values recorded for small molecules on this page: for cyclosporine in copovidone it lies between 1.6% and 5% peptide, and the paper gives neither a threshold nor a mechanism for the change [bapat2025b C5, C22, C45]. Congruence was asserted for systems where it was not computed: no normalized rate exists for any cyclosporine dispersion, and at 1.6% loading with SNAC the peptide in fact lagged (≈75.5% against ≈85% enhancer and ≈89.5% polymer at 30 min) [bapat2025b X2, R10.f]. The claim that a low peptide fraction removes the effect of peptide solubility holds for the decanoate ternaries but not for the SNAC ternaries [bapat2025b C41, X3]. The rates of the fastest systems rest on two or three points, because the compacts were nearly empty by the second sample [bapat2025b X8]. Several profiles plateau at ≈100–110% without a compact assay [bapat2025b X9]. Finally, congruent release of peptide and enhancer has not been shown to improve anything downstream: no permeability or absorption was measured [bapat2025b C6; fagan2025 C45]. In congruent dispersions the polymer still left more slowly than from a neat compact (2.35–2.76 against 3.49 mg/min/cm² for PVPVA; 0.22–0.29 of the neat rate for 10% ITZ), which the authors put "most likely" down to the drug's hydrophobicity or drug–polymer interactions [saboo2019 R8, C13; bhardwaj2018 R12, C49]. The link between congruency and droplets partly follows from how much drug had been released at the time of sampling [saboo2019 X14]. The neat PVPVA compact of the ledipasvir study weighed 250 mg against 100 mg for the dispersions [que2019 X13]. A congruency read-out over 5 min can miss later surface crystallization saboo2020 X7, and the criteria for the limit differ between papers [que2021 P9; hiew2022b X6]. For the miconazole dispersion on HP-50-Na the authors state polymer release "at the same normalized rate as the drug", but polymer percent release runs above drug release from 20 min (≈40.8 against ≈31.6%) [qi2022 C22, X4]. In the slowest HPMCAS-HF systems the drug left at about half the polymer's rate although the authors call release approximately congruent [bapat2025a X6, C13]. With SDS at 40% ritonavir release was already incongruent at 15 min, against the authors' "congruent" for the first 30 min yang2023 X4, C4.

← Concepts and formulas

concept · as used in Crystallization tendency

cosolvency

The change of a solute's solubility in a mixture of two solvents with their ratio; in this chapter water is an anti-solvent for hydrophobic drugs, and polymer plus sorbed water is treated as the drug's co-solvent inside a humid dispersion [marsac2008a C39, C26, C33; rumondor2009c C37].

Findings

F2, F10.

Related concepts

related to solid-liquid equilibrium solubility equation via activity coefficient; feeds supersaturation; related to Δχ effect.

Formulas

the log-linear cosolvency model of the 2008 paper is listed on the chapter page as table only.

How it is measured or obtained

Shake-flask solubility of the crystalline drug in a liquid analogue of the polymer (NMP for PVP), in water and in their mixtures at 25 °C, fitted with a log-linear cosolvency line; the water fraction of the polymer–water co-solvent is computed from sorption data marsac2008a P8, E5, E6.

Values in the library
materialvalue or classconditionspaper ids
nifedipine, felodipine in NMP–watermole fraction 0.05 and 0.10 at 0% water; ≈0.015 and ≈0.017 at ≈24 wt% water25 °Cmarsac2008a R12
nifedipine with 10 and 25% PVPco-solvent 23 and 18 wt% water75% RH, 25 °Cmarsac2008a R13
felodipine with 10 and 25% PVP≈16 wt% water75% RHmarsac2008a R13.c, R13.d
drug–PVP–water (argument)water–PVP "cosolvent" mixes less well with the drug as it takes up waterhumid dispersions[rumondor2009c C37, C44]
drugs in PEO compacts (outside)water lowers solubility in the water–PEO co-solvent but enlarges the solvent volume94% RHmarsac2008b C33

Evidence table: cosolvency.

Pitfalls and disagreements

The printed cosolvency fits do not reproduce their own points and cross at 16.5 rather than about 25 wt% water; the crossover rests on three water contents marsac2008a X2, X3. NMP stands in for PVP by assumption marsac2008a C33, X4.

← Concepts and formulas

concept · as used in Crystallization tendency

critical cooling rate

Rcrit (Rcrit,cool) is the slowest cooling of a melt that avoids detectable crystallization; the rate that avoids any nucleation, RNcrit, is faster still, and cooling between the two leaves nuclei in the glass baird2010 C50; baird2013 C2. It is the classical measure of glass-forming ability baird2010 C49.

Findings

F10, F11, F16.

Related concepts

measures glass-forming ability; estimated by Barandiaran-Colmenero method; contrasts with critical heating rate; related to quenched-in nuclei.

Formulas

Barandiaran-Colmenero critical cooling rate, Cabral critical cooling rate, critical cooling rate to avoid nucleation.

How it is measured or obtained

By cooling at progressively slower rates in a rapid heat–cool DSC until crystallization appears baird2013 P2, R5; by extrapolation with the Barandiaran–Colmenero or Cabral methods baird2010 P9; baird2013 P3; bounded by the fastest rate at which crystallization was still seen baird2013 R1, X6.

Values in the library
materialvalue or classconditionspaper ids
8 class I-A moleculesabove the fastest rate reached, 800–1100 °C/minrapid heat–cool DSCbaird2013 R1, X6
antipyrin, chlorpropamide, flufenamic acid, tolbutamide25 °C/minrapid heat–cool DSCbaird2013 R9
10 class II molecules5 °C/min or lessDSCbaird2013 R9
7 class I molecules (extrapolated)20 to 3.26 × 10⁴ K/minBC and Cabral, 5–20 °C/minbaird2010 R13
delamanidabove 50 °C/min (no glass at 50 °C/min)DSCduong2022a R3
Pitfalls and disagreements

Extrapolated values from narrow rate ranges are unreliable baird2010 C20. The authors advise that knowing Rcrit,cool exceeds 500 °C/min is "probably sufficient" to flag a difficult drug baird2013 C21. A low Rcrit,cool does not imply a stable glass, since reheating rates needed are far higher baird2013 C24.

← Concepts and formulas

concept · as used in Crystallization tendency

critical heating rate

Rcrit,heat, the slowest reheating of a glass through the undercooled melt that avoids crystallization baird2013 R9. For class I-B and II compounds it far exceeds the critical cooling rate, an asymmetry the authors report as a first for organic systems baird2013 C24, C26.

Findings

F10, F15, F16.

Related concepts

contrasts with critical cooling rate; measures glass stability; related to quenched-in nuclei and cold crystallization.

Formulas

none.

How it is measured or obtained

By reheating glasses at increasing rates in conventional (10–200 °C/min) and rapid heat–cool DSC (up to about 2000 °C/min), after cooling at 20 °C/min or at the maximum rate baird2013 P1, P2, P4.

Values in the library
materialvalue or classconditionspaper ids
bifonazole; celecoxib12.5; 15 °C/minafter 20 °C/min coolingbaird2013 R9.f, R9.g
tolazamide; droperidol35; 175 °C/minafter 20 °C/min coolingbaird2013 R9.n, R9.k
cinnarizine; clofoctol300; 400 °C/minafter 20 °C/min coolingbaird2013 R9.h, R9.i
dibucaine; acetaminophenabove 1000; above 1910 °C/minafter 20 °C/min coolingbaird2013 R9.j, R9.e
antipyrin, chlorpropamideabove 2000 °C/minclass I-B, after 20 °C/min coolingbaird2013 R9.a, R9.b
dibucaine after fast cooling300 °C/minafter 800 °C/min coolingbaird2013 R9.j, R11
Pitfalls and disagreements

The value depends on the prior cooling rate baird2013 C28; the Table 4 column heads of the source are misprinted and the instrument per value is not stated baird2013 X4, X9; heating rates above 1700 °C/min lie outside the stated calibration baird2013 X10.

← Concepts and formulas

concept · as used in Crystallization tendency

crystal growth inhibition

Reduction of the crystal growth rate by an additive, here dissolved PVPVA or HPMCAS, expressed as the effectiveness ratio E_G=R_{G,np}/R_{G,p}, the growth rate without polymer over the rate with it [moseson2020a E2; moseson2021 E1]. The mechanism proposed is surface poisoning: polymer globules adsorb on the crystal face and the adsorbed network "appears to limit the sites available for molecular integration" [moseson2020a C48, C50]. HPMCAS at nine times the drug concentration slowed seeded growth of posaconazole in FaSSIF V2 but did not stop it at 100 µg/mL drug: 10 µg/mL seeds left 43.5% at 30 min against 4.5% without polymer [shah2024 R22]. New aspect (Crystallization from solution): The slowing of the growth of drug crystals in a supersaturated aqueous solution by a dissolved additive, usually a polymer, expressed as the ratio of the initial growth rate without additive to that with it, E_g = R_{g0}/R_{gp}; above 1 the additive inhibits growth, below 1 it accelerates it [ilevbare2013b E5; ilevbare2012a E3, E4]. In these papers it is a kinetic effect at constant solubility, shown by solubility controls with the polymer [ilevbare2012c R1, C16]. It is the solution counterpart of the growth inhibition in supercooled liquids described below and is one of two steps of crystallization inhibition, the other being nucleation inhibition. Seeded growth ratios now include HPMCAS on felodipine (2.29 at pH 6.8, 1.28 at pH 3, and 2.7 to 12 depending on how the seeds were made) and bile salts on telaprevir seeds (E_G 1.0–4.6; complete stop with monomeric STCDC) [schram2015 R4; schram2016 R2; lu2017a R7, R10]. New aspect (Supersaturation from ASD dissolution): the 2008 felodipine study reads a flat concentration after an initial drop as growth inhibition: the drop takes the solution from the labile into the metastable zone ("it appears"), where only growth could lower the concentration further, so a plateau "strongly suggests" inhibited growth [konno2008a C27, C26]. The 2011 study reads slower growth of particles with HPMC than with PVP the same way [alonzo2011 C18]. For celecoxib, seeded desupersaturation with HPMCAS gave growth ratios R_0/R_p of about 70–158, less inhibition at the higher supersaturation, against an estimated nucleation ratio of ~4000, so both steps were slowed but nucleation far more ("it can be surmised") [xie2016a R12, R13, C37, C40]; the slow growth explains why crystals formed in dissolving PAA particles did not empty a medium holding HPMCAS [xie2016a C36]. In the supercooled liquid (formerly the page "crystal growth inhibition by polymers", from Crystallization tendency and Miscibility and drug-polymer interactions): the reduction of the crystal growth rate of a drug by a polymer dissolved in its supercooled liquid; the extent is thought to depend on specific drug–polymer interactions such as hydrogen bonding, and at higher levels on the polymer's effect on mobility trasi2014a C33; trasi2014b C19. The inhibition depends on the chemistry of the polymer at low levels, where it changes neither the Tg nor the melting point of the drug: at 3% w/w felodipine growth was slowed in the order PVP > PVP/VA > HPMCAS > PVAc kestur2010a C17, C32, C36. The factor is not a constant of the polymer: it grows as the temperature falls toward Tg and with polymer content kestur2010a C12, R6, R8, and the best polymer depends on the drug (PAA for bifonazole, PVP and PVP/VA for nimesulide at low temperature) kestur2011 C9, C10, C29. The authors tie the order to the strength or extent of drug–polymer hydrogen bonding and propose, as a "speculation", that crystal growth needs the drug to demix from the polymer at the growth front kestur2010a C40, C42.

Findings

ASD process: F25, F27, F29, F41. Crystallization from solution: F5, F6, F7, F8, F9, F10, F11, F12, F17, F18, F20, F23, F24, F27, F33. Supersaturation from ASD dissolution: F4, F5, F16, F22. Crystallization tendency: F19. Miscibility and drug-polymer interactions: F10, F11, F12, F13, F14, F17.

Related concepts

part of crystallization inhibition by polymers; acts on crystal seeding; slows desupersaturation; related to crystal growth rate; part of crystallization inhibition; contrasts with nucleation inhibition and with polymer effects on nucleation; related to polymer adsorption on crystal surfaces, step pinning and physical stability; contrasts with the polymer's effect on antiplasticization and thermodynamic driving force for crystallization; depends on hydrogen bonding and intramolecular hydrogen bonding.

Formulas

crystal growth inhibition effectiveness ratio, power-law crystal growth rate, initial growth rate from the desupersaturation slope, Kubota-Mullin model, overall mass crystal growth rate, relative nucleation rate from seeded and unseeded induction times, viscosity-limited normal growth law.

How it is measured or obtained

Seeded growth from a supersaturated solution at a stated supersaturation and polymer concentration, with the growth rate from the desupersaturation profile; adsorption by AFM in liquid (globule spacing) and crystal habit by SEM [moseson2020a P10, P15, P16; moseson2021 P12]. In the crystallization-from-solution papers: Seeded desupersaturation: a fixed mass of seed crystals (0.010 g in 50 mL for ritonavir, celecoxib and efavirenz; about 2.5 mg in 18 mL for felodipine) is placed in buffer with predissolved polymer, supersaturation is created by adding drug in methanol, and a fibre-optic UV probe on second-derivative spectra gives the slope of the first 2 min (5 min for felodipine) as the growth rate [ilevbare2013b P10; alonzo2012 P4]. Eg is the ratio of slopes, in triplicate [ilevbare2012c P12]. Rates can be normalized by seed mass and BET area ilevbare2013b E6. Unseeded controls and turbidity checks test that the slope is growth [ilevbare2012a R9; ilevbare2013b R5]. Rotating-disc seeds (felodipine) and seeds grown in situ in the vessel (telaprevir); the ratio is R_0/R [schram2015 P2; schram2016 P5; lu2017a P1, P2]. In the ASD dissolution papers: initial slopes of desupersaturation curves with equal crystal surface areas assumed (Eq. 5) [konno2008a E5, P6, C28]; particle size against time by light scattering [alonzo2011 P5]; seeded desupersaturation with 7 mg seeds in 50 mL, initial slope without polymer against the slope over 16 h with polymer [xie2016a P4, P11]. In the supercooled liquid: by comparing seeded growth rates of the neat drug and of drug with a low polymer level at the same temperatures trasi2014a P3; trasi2014b P2, with FTIR to assess drug–polymer hydrogen bonding trasi2014b P4. For the felodipine, bifonazole and nimesulide dispersions: a 10% w/w polymer dispersion made by rotary evaporation is diluted with crystalline drug in a cryogenic mill to 0.5–4.5% w/w, melted between cover glasses, nucleated over P2O5 at 40 °C (or at ambient temperature) and grown on a hot stage at 70–110 °C (or 18–100 °C), the slope of spherulite diameter against time being the rate kestur2010a P1, P2, P3, P4; kestur2011 P1, P2, P3. Whether the slope is a diameter or radius rate is not always stated, and the felodipine Fig. 3 slopes are about twice the Fig. 4 rates kestur2010a X3; kestur2011 X12. Hydrogen bonding is read separately by IR on films with 20–50 wt% polymer kestur2010a P6, C6.

Values in the library
materialvalue or classconditionspaper ids
IDM, PVPVAE_G 18 (5 µg/mL) and 56 (50 µg/mL polymer)50 µg/mL seeds, S = 6[moseson2020a R5]
BCL, PVPVA 70 µg/mL and HPMCAS 1 mg/mLE_G 8.6 and 158150 µg/mL, form 1 seeds[moseson2021 R5]
felodipine seeds, HPMCAS 5 µg/mL (outside)E_G about 2 to 12 depending on how the seed crystals were preparedS = 4[schram2016 R2, C13]
celecoxib, HPMCAS (outside)E_G about 130, 70 and 158 (R_0/R_p at 11/11, 22/5 and 22/22 µg/mL drug/polymer)11 and 22 µg/mL drug, pH 6.8, 37 °C[xie2016a R12]
ritonavir with cellulose polymers (outside)inhibition large at low supersaturation, marginal at the amorphous "solubility"seeded growth[ilevbare2012b C56]
felodipine in buffer, PVP, HPMC, HPMCAS (outside)initial slopes about 0.1399, 0.0398, 0.0044 and 0.0021 µg/mL·min⁻¹9.5 µg/mL start, 150 µg/mL polymer[konno2008a R16]
PCZ 100 µg/mL in FaSSIF V2, HPMCAS 0 / 3 / 9 times the drug; seeds 0 / 10 µg/mL% left at 30 min about 13 / 4.5; 66 / 10.5; 95 / 43.5seeded solution[shah2024 R22]
ritonavir, 34 polymersE_g 0.83–3.25; Pn-IPAAmd 3.25, CAP Adp 0.85 2.98, CAB Adp 0.81 2.70, HPMCAS about 2.545 µg/mL polymer, S 7.6, 37 °C[ilevbare2012c R9]
ritonavir, CA 320S Sub 0.90; 0.6312.6; 10.8sameilevbare2013b R9
celecoxib, HPMC; HPMCAS; PVPVA10.2; 8.5; 7.45 µg/mL polymer, S 6.6ilevbare2013b R9
efavirenz, PVPVA; CA 320S Sub 0.90about 4.9S 1.2ilevbare2013b R9
ritonavir, CAP Adp 0.856.40, 2.99, 1.56S 3.8, 7.6, 15.3[ilevbare2012b R5]
ritonavir, CAP Adp 0.85 + Pn-IPAAmdabout 12.1 (S 7.6); 3.3 (S 15.4)5 + 5 µg/mL[ilevbare2012a R11, R16]
felodipine, 1 µg/mL HPMCgrowth ratio 0.607S 10, 25 °C[alonzo2012 R5]
felodipine, HPMCAS-LF 5 µg/mLE_g 2.29 (pH 6.8), 1.28 (pH 3); 2.7 to 12 by seed route (pH 6.8)S 8 and S 4, 25 °C[schram2015 R4; schram2016 R2]
acetaminophen, 20 µg/mL polymerconcentration at 24 h 16.1 (HPMC), 15.8 (PVP), 13.4 (PAA), 13.0 (none) mg/mLS 1.5, 20 °Ctrasi2015 R11
telaprevir, 1.86 mM bile salts, δ 2.4E_G 3.2 (STC), 1.5, 1.4, 1.2; STCDC no desupersaturationpH 6.5, 37 °C[lu2017a R7, R10]
telaprevir seeds, 1:1 STDC–STCDC (binary-L, 1.86 mM; binary-H, 3.72 mM)E_G about 35.5 and 7.4, between STDC (3.8) and STCDC (>60)δ 7.9, 50 mM phosphate pH 6.5, 37 °C, first 10 min[lu2018 R5]
felodipine particles: none; 90:10 PVP dispersion; HPMC936 nm at 7 min; ≈218 → ≈459 nm (10–90 min); ≈280–307 nm at 90 min25 °C[alonzo2011 R10, R8]
felodipine particles at 37 °C, PVP against HPMC dispersions≈441 nm by 20 min against ≈287–307 nm37 °C[alonzo2011 R11]
paclitaxel seeds with 20 µg/mL HPMCASgrowth continued at S = 88 and left ≈7.9–25.3 µg/mL after 16 h; seeds grew dendritic fingers ≈10.5–16.8 nm widepH 6.8, 37 °C[que2018 R14, R12]
praziquantel with HPMCAS-MF against HPMC E5 LVno concentration drop with HPMCAS outside fed media; HPMC lost dissolved drug at 1.5 mg/mL37 °C[alvarenga2025 R14, C28]
five drugs with 10 mol% PVPgrowth reduced about 3- to 14-fold (authors: "around 5−10"), one point about 50-foldsupercooled liquid, 40–120 °Ctrasi2014a R6, X6, C10
nilutamide≈5 (alone), ≈3.8 (PAA), ≈1.1 µm/s (PVP)supercooled liquid, 80 °Ctrasi2014b R3
felodipine with PVP K-29/322- and 6-fold at 110 °C, 3- and 35-fold at 70 °C (1.5 and 4.5 wt%)supercooled liquid, meltkestur2010b R13
bifonazole with PAA5-fold at 100 °C, 15-fold at 18 °Csupercooled liquid, 5 wt%kestur2011 R7
flutamide with PVPabout one order of magnitudesupercooled liquid, 10 mol%trasi2012b R3
celecoxib with PVP; with TPGSmore than one order slower; about two orders faster at 70 °Csupercooled liquid, 10% w/wmosqueragiraldo2014 R1, C9
felodipine with 3% PVP, PVP/VA, HPMCAS, PVAcrate ratios at 70 °C ≈10.0, 6.5, 4.4, 1.2; at 110 °C ≈2.9, 1.1, 1.1, 1.1supercooled liquid, form II, hot stage, triplicatekestur2010a R6
felodipine, 4.5% w/w at 90 °C≈4.7 × 10⁻⁹ (PVP), 6.4 × 10⁻⁹ (PVP/VA), 1.7 × 10⁻⁸ (HPMCAS), 3.8 × 10⁻⁸ m/s (PVAc) against 6.7 × 10⁻⁸ m/ssupercooled liquid, log-linear in polymer contentkestur2010a R8
bifonazole with 5% PAA; PVP; HPMCASratios ≈15, 5.9, 4.0 at 18 °C; ≈5.1, 2.1, 1.7 at 100 °Csupercooled liquid, melt-quenched filmskestur2011 R7
nimesulide with 5% PVP; PVP/VA; PVAcratios ≈21, 16, 3.4 at 18 °C; ≈4.7, 3.1, 2.0 at 80 °Csupercooled liquid, melt-quenched filmskestur2011 R9
Pitfalls and disagreements

E_G falls as supersaturation rises and depends on seed preparation, polymer concentration and polymer chemistry, so values from different systems are not comparable [ilevbare2012b C56; schram2016 C13; moseson2020a X12]. Adsorption was imaged on bulk crystals only, not on residual crystals [moseson2020a X5]. The threshold for "effective" is applied loosely: Eg above 1 is the stated rule, but counts of effective polymers match a cut-off near 1.2, and polymers at 1.0–1.4 are called ineffective [ilevbare2013b X3; ilevbare2012c X7]. Eg depends on supersaturation, polymer concentration, pH and buffer strength, so ratios from different papers are comparable only roughly; absolute rates cannot be compared because seed mass, volume and methanol fraction are often not printed [ilevbare2012c X16; ilevbare2012b X12]. Most 2013 ritonavir values were reported before ilevbare2013b X15. A polymer that raises the solubility lowers S and so flatters Eg [ilevbare2013c X7; ilevbare2012a X6]. The ratio depends on the seed route and the dose [schram2016 R2]; rates are not normalised to area in the felodipine and telaprevir papers [schram2016 X12; lu2017a C5]. For a bile salt mixture the "intermediate" ratio rests on an off-scale value for STCDC and a wide spread for the mixture (about 16.7–54.5), and secondary nucleation at δ 7.9 was assumed small [lu2018 X7, C8]. Polymers that form globules on an amorphous drug surface "may" need a higher concentration to block growth, which was not measured [mosqueragiraldo2018a C41]. No precipitate was examined in the 2008 felodipine study and surface areas were assumed equal [konno2008a X4, X7]. Seeded and unseeded tests of felodipine with HPMC later found growth at 0.607 of the polymer-free rate but an estimated nucleation-rate ratio of 7.05 × 10⁻⁴, placing the main effect on nucleation [alonzo2012 R5, R17]. For nifedipine polymorph conversion the polymers are read as inhibiting α nucleation, not growth raina2014b C32. The celecoxib slopes are not printed and R_p was taken over 16 h while the plot covers 4 h, so the ratios cannot be reproduced; seeded growth was measured with HPMCAS only, not HPMC [xie2016a X8, X5]. A polymer that blocks nucleation need not stop growth: HPMCAS held unseeded paclitaxel at S = 88 for more than 16 h but only slowed growth on seeds [que2018 R3, C30]. In the supercooled liquid: inhibition of growth and of nucleation need not coincide either: for acetaminophen, HPMCAS inhibited nucleation best but growth least trasi2012a C46. Some additives accelerate growth sato2016 R11. The molar basis of the PVP level was not defined trasi2014a X7. The text of the felodipine paper overstates two points against its own figures: the 105 → 70 °C fall is about 51-fold, not 3 orders of magnitude, and PVP still inhibits about 3-fold at 110 °C where the polymers "appear to have no effect" kestur2010a X1, X2. The nimesulide text gives 2–10-fold, the plot up to about 21-fold kestur2011 X7. The hydrogen-bond order was measured at 20–50 wt% polymer and one temperature, and bifonazole–polymer bonding was not measured kestur2010a X5; kestur2011 X9. The melting-point argument used form I in physical mixtures while form II grows kestur2010a X7. Only growth was measured, not nucleation kestur2010a X10. The two media are explained differently: in solution the authors propose surface poisoning by adsorbed polymer [moseson2020a C48, C50], in the supercooled liquid drug–polymer hydrogen bonding and mobility, with demixing at the growth front as a "speculation" trasi2014a C33; kestur2010a C40, C42, so ratios from the two media are not interchangeable (synthesis: different media and proposed mechanisms).

← Concepts and formulas

concept · as used in Crystallization tendency

crystal growth rate

The speed at which a crystal front advances into the surrounding supercooled liquid or glass, taken as the slope of crystal size (length, radius or diameter) against time trasi2014a P1; trasi2014b E1. Its temperature dependence is bell-shaped, with a maximum crystal growth velocity between Tg and Tm trasi2014a C22. This draft comes from the crystallization-tendency chapter (growth from melts); growth from supersaturated solution, measured by desupersaturation, is covered by other chapters ilevbare2013b R7. New aspects (Miscibility and drug-polymer interactions): Growth of felodipine from the supercooled liquid is expected to depend on a thermodynamic part (the free energy difference between liquid and crystal) and a kinetic part (viscosity), as in u = (k/\eta)[1 - \exp(-\Delta G/RT)], and also on molecular recognition at the crystal surface kestur2010a C30, E1. The growth rate of felodipine peaks at about 105 °C and a polymer at 3% shifts the apparent maximum to 110 °C (not for PVAc) kestur2010a R5, C11. The polymer effect, expressed as the ratio of growth rates without and with polymer, is not constant but grows toward the glass transition kestur2010a R6; kestur2011 C8, C10. New aspect (Crystallization from solution): growth from supersaturated aqueous solution is measured as the initial bulk growth rate of seed crystals, the slope of the dissolved drug amount against time over the first 2 min (5 min for felodipine), in µg/mL/min or µg/min, and normalized by seed mass and BET area (mg min⁻¹ m⁻²) to compare drugs [ilevbare2013b E6, E8; alonzo2012 P4]. It follows the empirical law R_G = k_G S^{g} with order g between 1 (diffusion control) and 2 (integration control) [alonzo2012 C3, E2]. New aspect (Physical stability): in the physical-stability papers the growth rate is the slope of crystal diameter (melt films) or radius (thin films at fixed RH) against time, of existing crystals nucleated first, so that growth is measured apart from nucleation kestur2010b P3; rumondor2010 P1, P5, C7. New aspect (Supersaturation from ASD dissolution): in the ASD dissolution papers a growth rate from solution is the mass gained by crystals per unit surface area and time, R_G = \frac{1}{A_c}\frac{dM_c}{dt}; with equal surface areas assumed, the slope of the falling concentration ranks growth rates [konno2008a E5, C28].

Findings

Crystallization tendency: F1, F2, F3, F5, F6, F7, F15, F19, F21. Miscibility and drug-polymer interactions: F10, F11, F12, F13. Crystallization from solution: F3, F5, F7, F9, F10, F23, F24, F33, F42. Physical stability: F3, F9, F11, F15, F17, F18, F19, F21, F22, F27, F28, F29, F35. Supersaturation from ASD dissolution: F5.

Related concepts

part of crystal growth; includes maximum crystal growth velocity; depends on melt viscosity and thermodynamic driving force for crystallization; contrasts with nucleation; related to crystal growth inhibition by polymers and surface crystallization; includes spherulitic growth; depends on supercooled liquid; lowered by crystal growth inhibition by polymers; inhibited in crystal growth inhibition; limited at high supersaturation by kinetic roughening; related to integration-controlled crystal growth. Regime diffusion-controlled crystal growth; modified by crystallization inhibition by polymers and moisture-induced crystallization; related to surface-enhanced crystal growth; faster mode near Tg: glass-crystal growth mode; split from nucleation and crystal growth; related to desupersaturation.

Formulas

temperature of maximum crystal growth rate, Jackson interface-kinetics growth rate, linear fit of melt viscosity on Tm − Tg, linear fit of normalized growth rate on melting entropy; viscosity-limited normal growth law, power-law crystal growth rate, initial growth rate from the desupersaturation slope; empirical log-linear growth-rate equation, linear crystal growth rate from size-time slope, crystal growth inhibition effectiveness ratio, overall mass crystal growth rate.

How it is measured or obtained

Hot-stage polarized light microscopy of melt-quenched films seeded at the edge with the form of interest, triplicate where stated trasi2014a P1; trasi2014b P3; with the top coverslip removed for surface growth trasi2014a P2; from the crystallized area when the front is irregular trasi2014b P3; the size–time plot is linear trasi2012a R4; kestur2011 R4. In the felodipine, bifonazole and nimesulide studies: the slope of spherulite diameter against time on a hot stage at 70–110 °C (felodipine) or 18–100 °C (bifonazole, nimesulide), triplicate; slower growth below 40 °C was followed in ovens or coolers with periodic pictures kestur2010a P4; kestur2011 P3. Whether the slope is a diameter or a radius rate is not always clear: the felodipine Fig. 3 slopes are about twice the Fig. 4 rates kestur2010a X3; kestur2011 X12. From solution, by seeded desupersaturation with a fibre-optic UV probe on second-derivative spectra, in triplicate (F5) [ilevbare2013b P10; ilevbare2012a P8]. In the physical-stability papers: melt-quenched felodipine between cover glasses, nuclei formed at 40 °C, grown on a hot stage at 70–110 °C, with crystals 5.5 ± 2 µm thick so that diameter slopes can be compared kestur2010b P2, P3, R19, C5. Uncovered spin-coated films, spherulites nucleated on a 90 °C hot plate, radius read from photomicrographs at four or more times for three or more crystals at each RH rumondor2010 P1, P5. Seeded edge growth of melt-quenched films of flutamide, acetaminophen and celecoxib, triplicate where stated; the flutamide paper reports diameter, the acetaminophen paper radial growth rates trasi2012b P2, R3; trasi2012a P2, R6; mosqueragiraldo2014 P3. Felodipine form I at a free surface or seeded in the bulk, form II unseeded between two cover glasses kestur2013a P2, P3. In the ASD dissolution papers: the initial linear slope of solvent-shift desupersaturation curves (felodipine from 9.5 µg/mL with 150 µg/mL polymer) [konno2008a P6].

Values in the library
materialvalue or classconditionspaper ids
nimesulideMCV 216 µm/s, the fastest of 31 drugsbulk melttrasi2014a R1.a
clotrimazoleMCV 0.02 µm/s, the slowestbulk melttrasi2014a R1.ae
vemurafenib α-needles; β≈106 µm/s at 250 °C; ≈110 µm/s at 230 °Csupercooled liquidlu2016 R12.o, R12.m
nilutamide stable form≈0.00015 µm/s at 30 °C to ≈40 µm/s at 100 °Cseeded filmstrasi2014b R8
felodipinemaximum near 105 °C, 3 orders lower at 70 °Cmelt filmskestur2010a R5, C10
felodipine form Iabout 6 times faster at the surface than in the bulk30 °C, 0% RHkestur2013b R12
felodipine filmsabout 2-fold rise over 13–80% RH (≈0.030 → 0.066 mm/min), 40-fold at 87% RH (2.3–2.8 mm/min at 87–93% RH)25 °Crumondor2010 R1
bifonazoleabout 5.5 orders lower from 100 to 18 °Cmelt-quenched filmskestur2011 R7
flutamidenearly 2 orders lower from 70 to 40 °Cfilmstrasi2012b R3
acetaminophennearly 2 orders lower from 110 to 70 °Cfilmstrasi2012a R6
indomethacin γ; α; δMCV 4.07 × 10⁻⁷; 4.34 × 10⁻⁶; ≈1.5 × 10⁻⁷ m/s, at 135, 135 and 110 °Cpure melttian2017 R7, R8, R9, R10
celecoxib with TPGSabout two orders faster than celecoxib alone at 70 °C10% w/wmosqueragiraldo2014 C9
felodipine form II≈3.5 × 10⁻⁹ (70 °C), ≈6.9 × 10⁻⁸ (90 °C), ≈1.8 × 10⁻⁷ m/s (105 °C)neat, hot stagekestur2010a R5.a
felodipine, 3% PVP; PVP/VA; HPMCAS; PVAcat 70 °C ≈3.5 × 10⁻¹⁰, 5.4 × 10⁻¹⁰, 7.9 × 10⁻¹⁰, 3.0 × 10⁻⁹ m/sform IIkestur2010a R5.e, R5.d, R5.c, R5.b
bifonazole neat; with 5% PAA≈1.7 × 10⁻¹² and ≈1.1 × 10⁻¹³ m/s at 18 °C; ≈4.7 × 10⁻⁶ and ≈9.2 × 10⁻⁷ m/s at 100 °Cmelt-quenched filmskestur2011 R7.a, R7.i
nimesulide neat; with 5% PVP≈3.7 × 10⁻¹¹ and ≈1.8 × 10⁻¹² m/s at 18 °C; ≈8.5 × 10⁻⁶ and ≈1.8 × 10⁻⁶ m/s at 80 °Cmelt-quenched filmskestur2011 R9.a, R9.g
ritonavir; celecoxib; efavirenznormalized 0.20; 0.91; 2.97 mg min⁻¹ m⁻² at S 1.2–1.3seeded, 37 °Cilevbare2013b R7
celecoxib; ritonavir at S 6.6; 7.66.23; 6.14 mg min⁻¹ m⁻²sameilevbare2013b R7
celecoxib; ritonavir at S 14.7; 15.419.08; 14.32 mg min⁻¹ m⁻² (217.5 and 181.5 µg min⁻¹)sameilevbare2013b R7
ritonavir without additive, 10 µg/mL1.56 µg mL⁻¹ min⁻¹ (text), about 1.4 (figure)100 mM buffer, S 7.6[ilevbare2012a R10, X5]
ritonavir without additive, S 3.8; 7.6; 15.4about 0.72; 1.4; 3.63 µg mL⁻¹ min⁻¹same[ilevbare2012a R15]
felodipine without HPMC, S 3 to 12about 0.078 to 0.62 µg mL⁻¹ min⁻¹; g about 1.5025 °C[alonzo2012 R4, R13]
telaprevir seeds, no bile salt5.4, 58.6, 89.2 µg/min at δ 2.4, 7.9, 12.5 (per 50 mL vessel)pH 6.5, 37 °C[lu2017a R2, X12]
felodipine seeds, no polymerconcentration 4.0 → 1.7 µg/mL in 50 minS 8, 25 °C, 1000 rpm[schram2015 R1]
atazanavir, posaconazole, six mediaSHG size growth about 2–55 (atazanavir) and 20–156 nm/min (posaconazole); highest in blank buffers37 °C, relative per field[elkhabaz2021 R15, X13]
felodipine form II (PVP grade study)3.5 × 10⁻⁹ m/s (70 °C) to 1.7 × 10⁻⁷ m/s (105 °C, maximum)melt filmskestur2010b R7, C9
felodipine with 3 wt% PVP K-12, K-29/32, K-9011-, 15-, 21-fold slower at 70 °C; 3-, 3-, 6-fold at 110 °Cmelt filmskestur2010b R8
felodipine with 0.5–4.5 wt% PVP K-29/32ln rate linear in concentration; slope −0.82 (70 °C) to −0.49 per wt% (105 °C)melt filmskestur2010b R15, E1
felodipine with 10% PVP vs 10% HPMCAS5.8 × 10⁻⁵ vs 1.3 × 10⁻³ mm/min (13% RH); 7.3 × 10⁻² vs 2.4 × 10⁻³ (93% RH)room temperaturerumondor2010 R5, R3
flutamide with 10 mol% PVP≈0.13 against ≈1.4 µm/s at 40 °Cmelt films, diameter ratetrasi2012b R3.a, R3.h
acetaminophen with 10% w/w PAA≈0.43 against ≈5.3 µm/s at 80 °Cmelt films, radial ratetrasi2012a R6.a, R6.h
celecoxib≈0.002 (70 °C) to ≈12 µm/s (120 °C); 19–38-fold slower with 10% PVPseeded melt filmsmosqueragiraldo2014 R1.a, R1.b
felodipine form I against form II, bulkform I faster at 40–100 °C: ≈4700-fold at 40 °C, 7–9-fold at 80–100 °Cmelt filmskestur2013a R8, X5
felodipine form I with 3% PVPsurface 1.45 × 10⁻⁹, bulk 6.37 × 10⁻¹¹ m/s (ratio 22.8)30 °C, 0% RHkestur2013b R12
ATP(S), ATP(R)0.022–0.032 µm/s (40 °C) to 21–34 µm/s (90 °C); S 1.2–1.6 times fastermelt films[sato2015 R5.a, R5.b, X2]
griseofulvin with 5% PHB≈0.015 → ≈0.15 µm/s at 120 °C (diameter rate)melt filmssato2016 R4.a, R4.c
celecoxib, alone and with 10% PVP K29/32≈0.010 and ≈0.00003 µm/s at 80 °Cmelt films[xie2016b R21]
felodipine, buffer / PVP / HPMC / HPMCASdMc/dt ~0.1399 / ~0.0398 / ~0.0044 / ~0.0021 µg/mL-minpH 6.8, 37 °C[konno2008a R16]
Pitfalls and disagreements

The MCVs of the 31-drug set have no stated derivation and some disagree with the plotted rates; no replicates are given trasi2014a X3, X17. Rankings change with the temperature scale (T/Tg against Tm − T) trasi2014a C7 and between surface and bulk trasi2014a C8. Rates above Tg cannot readily be extrapolated to storage temperatures trasi2014a C35, and surface-enhanced or sudden growth modes near Tg can make high-temperature data overpredict stability kestur2013a C7. Near Tg, diffusion decouples from viscosity trasi2014a C21. The text says the felodipine rate falls by 3 orders of magnitude from 105 to 70 °C (the figure in the table above), the plot gives about 51-fold kestur2010a X1. The text says the polymers "appear to have no effect" at 110 °C, but PVP still gives a ratio of about 2.9 kestur2010a X2. All rates of the felodipine, bifonazole and nimesulide studies are plot readings from log axes and the error bars are not defined kestur2010a X9; kestur2011 X6. The bifonazole orders of magnitude in the text (5.5, as in the table above) differ from the plot kestur2011 X6. In solution, the same no-polymer condition gives 71.0, 69.80 and about 68 µg min⁻¹ for celecoxib in three places of one paper ilevbare2013b X7. Seed amount, volume and methanol fraction are often not printed, so absolute solution growth rates cannot be compared across studies [ilevbare2012c X16]. Efavirenz shows secondary nucleation above 10 µg/mL, which confounds its seeded rate ilevbare2013b R5. The fall of pure felodipine growth from 105 to 70 °C is about 49-fold in the PVP grade paper (synthesis: 1.7 × 10⁻⁷/3.5 × 10⁻⁹ kestur2010b R7) but 3 orders of magnitude in the text of the other 2010 paper kestur2010a C10. The melt growth model takes the diameter slope as the radial front speed kestur2010b X5. Replicates and error-bar meaning are not stated for the melt study or for the polymer films at fixed RH kestur2010b X17; rumondor2010 X7. Diameter and radial growth rates differ by a factor of two, so flutamide and acetaminophen rates cannot be compared as printed trasi2012b R3; trasi2012a R6. The measured felodipine front changes habit with temperature kestur2013a X12. The PHB paper reports diameter rates, twice the radial rate sato2016 X8. Table 2 of the indomethacin polymorph paper interchanges the δ and γ maxima of the pure drug tian2017 X1. Two celecoxib papers differ about tenfold in the factor by which 10% PVP slows growth [mosqueragiraldo2014 R1; xie2016b R21]. The slopes come from one panel without uncertainty, from curves that start at different 5-min levels, and no surface area was measured [konno2008a X7]. The buffer slope reproduces from the plotted points (≈0.14 µg/mL-min) [konno2008a E5].

← Concepts and formulas

concept · as used in Crystallization tendency

crystal growth

The second step of crystallization: existing nuclei or seeds grow by molecules crossing the crystal–liquid interface and attaching to growth sites trasi2014a C19; tian2017 C22. The usual growth models (normal, screw dislocation, two-dimensional surface nucleation) differ in the interface structure but all contain a mobility term and a driving-force term trasi2014a C19. Because nuclei are detected only after they grow, a measured induction time is the sum of nucleation and growth times chen2015 E1, C5. New aspect (Bioperformance): the growth rate is written as a power law in supersaturation, R_G = k_G A (C_b - C^*)^g, and growth rates of ezetimibe were taken from SHG [purohit2018 E3; elkhabaz2017 R23]. Growth on tacrolimus seeds was fitted with the same power law, dC/dt = k_g A (C - C^*)^n, with n = 1.677 (R² 0.9896), close to the n of about 2 expected for integration-controlled growth with a polymer adsorbed on the crystal [hate2019 E4, E5, R18, C44]. New aspect (Supersaturation from ASD dissolution): growth is the second step that empties a supersaturated solution after nucleation; the two-route scheme of dispersion dissolution writes it as proportional to the excess concentration, R_G \propto (C - C_{eq}) (cited) alonzo2010 E5, C43. A celecoxib study states that crystals formed in the wetted matrix act as seeds that grow quickly and limit the supersaturation, and reads the induction time as nucleation plus growth to a detectable size xie2015 C31, E1.

Findings

Crystallization tendency: F1, F19. Bioperformance: F10, F12, F16, F29. Supersaturation from ASD dissolution: F5, F9, F16, F22.

Related concepts

measured by crystal growth rate; contrasts with nucleation; related to crystal growth inhibition by polymers and surface crystallization. Seeded by crystal seeding; inhibited by crystal growth inhibition.

Formulas

Jackson interface-kinetics growth rate. Power-law crystal growth rate, overall mass crystal growth rate.

How it is measured or obtained

As a growth rate (see crystal growth rate); by the morphology of the growing crystals lu2016 R7; in solution by desupersaturation after seeding xie2015 C31. It is inferred from the fall of concentration in non-sink tests [purohit2018 C43] and from the SHG mean size over time [elkhabaz2017 P11]. In dissolving dispersions and in methanol-shift solutions: size of submicron particles against time by light scattering [alonzo2011 P5]; desupersaturation slopes [konno2008a P6].

Values in the library
materialvalue or classconditionspaper ids
bifonazole, nimesulidespherulite size linear in timemelt filmskestur2011 R4
acetaminophen with polymersspherulites; radius linear in time, R^2 > 0.99melt filmstrasi2012a R4
vemurafenibfour forms in six morphologies (needles, spherulites, feathers, clusters)supercooled liquidlu2016 R7
nilutamide stable formgrowth front smooth at 70 °C, rough at 60 °C, with a 50-fold rate dropseeded filmstrasi2014b R4, C7
flutamide dispersion particlesnumber and size of needle crystals increase with storage40 °C/75% RHcorreasoto2017 R2, C12
ezetimibe ASDsgrowth rate 0.020–1.0 µm/min (as printed)buffer, FeSSIF, Ensure Plus[elkhabaz2017 R23]
tacrolimusgrowth rate rises steeply with supersaturation; 450 mL gives twice the supersaturation of 900 mLUSP medium I[purohit2018 C52, C55]
FTM ASD (SHG)spots at 8 days, dense field at 60 days75% RHcorreasoto2017 R2, R4
tacrolimus on seedsn 1.677; growth constant 1.17 × 10⁻⁴ (printed unit cm/s, unclear); absorption leads below S about 18.5 and growth above ithollow-fibre module, seeds from the crystallized generic[hate2019 R18, R19, X3, X16]
felodipine particles from a 90:10 HPMC dispersion≈218 nm at 5 min, ≈337 at 90 min, ≈720 nm near 10 h, then settling25 °C[alonzo2011 R6]
felodipine particles, dispersion against methanol shift with HPMC≈183–203 → ≈280–307 nm in 90 min, alike25 °C[alonzo2011 R8]
same at 37 °Cdispersion-born ≈259–274 nm against ≈194–219 nm at 5–10 min37 °C[alonzo2011 R11]
celecoxib seeds, 22 and 11 µg/mL, no polymerno lag; ≈22.5 → ≈4.0 and ≈12.8 → ≈3.7 µg/mL in 30 minpH 6.8, 37 °C[xie2016a R11]
paclitaxel with 18 wt% seeds, no polymerfrom 31.6 µg/mL to ≈12–13 µg/mL in 2 min with sonicated seeds against ≈21.6 and ≈26.1 µg/mL with long needlespH 6.8, 37 °C[que2018 R7]
Pitfalls and disagreements

Near Tg some crystals switch to diffusionless growth and diffusion decouples from viscosity trasi2014a C21. A change in growth morphology can change the rate abruptly trasi2014b C7. Once crystallization starts it is expected to continue, so early detection matters correasoto2017 C4. In the SHG study of ezetimibe the nucleation-rate unit is implausible and growth rates are relative [elkhabaz2017 X7]. In the tacrolimus seed study the unit of the printed growth constant does not fit the equation, the x-axis of the log-log fit is log(C − C*) rather than log(S − 1), and the crossover near S 18.5 assumes a constant crystal area (F29) [hate2019 X1, X2, X3, X16]. The crystallinity of the early felodipine particles was inferred, not measured by diffraction [alonzo2011 X7]; the match of dispersion-born and methanol-born particles holds at 25 °C, not at 37 °C [alonzo2011 X19]. Per unit mass the growth area, not the seed mass, set the loss: sonication raised the needle-end area about 7–9-fold [que2018 R6, C5].

← Concepts and formulas

concept · as used in Crystallization tendency

crystal seeding

Crystals already present in a dispersion or added to the medium act as templates for growth and consume supersaturation; the relevant property is the growth area and interface structure of the seeds, not their mass [que2018 C5]. In the ASD process papers, residual crystals from hot melt extrusion are seeds that are small, embedded in the amorphous solid and possibly defective, and are "routinely" more potent than bulk seeds [moseson2020a C55; moseson2021 C65, C66, C74]. In the posaconazole–HPMCAS pH-shift tests residual crystals of 0.3–3.2% act as seeds: supersaturation ended earlier, by growth of the crystals present and by secondary nucleation, both inferred from solution concentration and particle counts [shah2024 C45, C47, R14]. New aspect (Bioperformance): crystals already present in an amorphous product on which dissolved drug can grow without nucleation; their growth removes supersaturation [purohit2018 C40, C43]. Seeds were taken from the maximally crystallized generic tacrolimus capsule and added to solutions at 20 and 50 µg/mL; crystals that formed inside the product were smaller than purchased reference crystals of the same form [hate2019 R17; purohit2019 R6, R8, C56]. New aspect (Supersaturation from ASD dissolution): a dispersion can seed itself. Crystals that form in the wetted, undissolved matrix during a gastric stage are released when the polymer dissolves at intestinal pH and then act as seeds for the supersaturated solution; for posaconazole the authors state that "These seed crystals were transferred to the intestinal compartment and promoted further crystallization" [elkhabaz2019a C56], and for delamanid that surface crystals "can act as seeds" while release and crystallization compete [nguyen2023c C44, C49].

Findings

ASD process: F26, F28, F29, F41. Bioperformance: F16, F28, F29, F30. Supersaturation from ASD dissolution: F22, F24, F25, F26, F27.

Related concepts

produced by residual crystallinity; read through desupersaturation; slowed by crystal growth inhibition; influenced by crystal defects. Related to crystal growth, secondary nucleation and endogenous crystallization; produced by matrix crystallization and surface crystallization; acts through seeded crystal growth; avoided by enteric polymer coating of the tablet.

Formulas

power-law crystal growth rate.

How it is measured or obtained

Seeded desupersaturation: bulk crystals (D50 21.9 µm) added at 10, 25 or 50 µg/mL to 50 µg/mL indomethacin, with the growth rate in µg/s [moseson2020a P10, R4]; external form 1 seeds equal to 10% of a bicalutamide dose added to a crystal-free dispersion and read by Raman as a relative growth rate [moseson2021 P13, P17, R22]. Potency of residual seeds is read by comparing extrudates with seeded crystal-free dispersions at the same amorphous dose [moseson2020a P13, P14]. Seeded desupersaturation of posaconazole in FaSSIF V2: 20 or 100 µg/mL drug, HPMCAS at 0–9 times the drug, seeds of 0–10 µg/mL, percent left at 30 min [shah2024 P16, R22]. In the tacrolimus capsules crystals were read by XRPD (limit of detection 10%) and inferred from desupersaturation in non-sink dissolution [purohit2018 R2, R7]. For self-seeding in pH-shift tests: crystals are detected in the acid stage by SHG, which gives signal only from crystals, beside two-photon excited fluorescence of all drug, and their ratio estimates the share of the undissolved drug that is crystalline [elkhabaz2019a P10, P11, E1]; on dispersion films and powders by polarized light and SEM after 1 h in acid, with PXRD of the powders [nguyen2023c P12, P13, P14]. The effect is read from the release after the pH shift against a run without an acid stage [elkhabaz2019a R6, R7].

Values in the library
materialvalue or classconditionspaper ids
IDM bulk seeds, no polymergrowth rate about 0.33, 0.71, 0.99 µg/s at 10, 25, 50 µg/mL seedsS = 6, pH 4.5[moseson2020a R4.a]
IDM–PVPVA, 50 µg/mL amorphous dosebulk seeds up to 100% of the dose: C720 about 43.2 → 39.8 µg/mL; extrudates with 3–25% crystals: about 27.0–37.3 µg/mL12 h[moseson2020a R9, R10]
BCL–PVPVArelative form 1 growth rate about 0.088 (M/155, 3.5% crystals), about 0.071 (S/155, below 1%), about 0.044 (10% external seeds)Raman slurry[moseson2021 R22]
paclitaxel–HPMCAS 49/51 (outside)residual concentration about 25 (bulk API) to about 8–10 µg/mL for other seed types; seed mass not the relevant parameter18 wt% seeds, 16 h[que2018 R14, C5]
fresh generic tacrolimus capsules (outside)seeds below 5% of the drug cause desupersaturation; growth slow at first because the surface area is smallnon-sink, 100 mL[purohit2018 C45, C46]
PCZ 100 µg/mL in FaSSIF V2, seeds 10 µg/mL4.5% left at 30 min without polymer; 10.5% with HPMCAS at 3 times the drug; 43.5% at 9 timesseeded solution[shah2024 R22]
PCZ–HPMCAS 10% DL with 0.3% crystalssupersaturation down to about 18.5 µg/mL by 150 min against about 125 µg/mL at 300 min without crystalspH shift at 60 min[shah2024 R12, R14]
tacrolimus generic capsulesfresh seeds below 5% of the drug (inferred); peak 98% then 39.5% in 100 mLUSP medium I[purohit2018 C46, R7]
felodipine, HPMCnucleation inhibition counts far more than growth inhibitionseeded and unseeded[alonzo2012 C40]
posaconazoleseeded desupersaturation faster with more seeds and more drug, slower with more HPMCASFaSSIF V2[shah2024 C37, R22]
tacrolimus, seeded solutionsdesupersaturation rate about 0.39 at 50 µg/mL and 0.075 at 20 µg/mLpolymer in solution, hollow fibre[hate2019 R17]
tacrolimus crystals in the generic capsule1–5 µm against 5–25 µm for reference monohydrate crystals; same formSEM, after 5 weeks at 40 °C/75% RH[purohit2019 R6, R8]
posaconazole–HPMCAS 10 / 25 / 50% drugestimated matrix crystallization ≈0.026 / ≈0.050 / ≈68% of the undissolved drug30 min in FaSSGF, 37 °C[elkhabaz2019a R20]
the same, after transfer10% and 25% held ≈22–24 and ≈21–22 µg/mL for about 150 and 60 min; 50% fell to ≈6 µg/mL within 30 minFaSSIF pH 6.5[elkhabaz2019a R6, R7, R8]
the same, without an acid stage≈21 µg/mL at 180 min (10%) and ≈22–23 µg/mL to 60 min (25%)FaSSIF only[elkhabaz2019a R6, R7]
delamanid salt dispersions, particles after the pH shiftcovered with small crystals within 30 minpH 6.5[nguyen2023c R21]
paclitaxel–HPMCAS dispersion with 18 wt% added seedsthe amorphous solubility was still reached, then lost over hourspH 6.8, 37 °C[que2018 R13]
Pitfalls and disagreements

Surface area and defects of residual crystals were not measured, so the extra potency is an inference [moseson2020a X9; moseson2021 X10]. Bulk seeds at a larger mass can grow more slowly than a small mass of embedded residual crystals; micron-sized seeds would "still not approximate" crystals below 100 nm [moseson2021 R22; moseson2020a C42]. Seed potency also depends on supersaturation and on the polymer [moseson2020a C54, C56]. The focused beam reflectance signature read as secondary nucleation for the crystal-containing 25% dispersion has the same form as the one read as primary nucleation for the crystal-free one, and the separate rates of growth and nucleation were not measured [shah2024 K3, K4, X14, X13]. Added seeds may not mimic crystals formed inside the ASD [purohit2018 C7]. Growth in the tacrolimus tests was inferred from concentration [purohit2018 X9]. Seeds did not always cause crystallization: at 240 and 100 mL the lower maximum supersaturation and the membrane loss kept growth small, at 40 mL it was large (F28, F29) [hate2019 C46, C49]. The rate units printed with the seeded runs are inconsistent with the profiles [hate2019 X1]. No seeding experiment was run in either pH-shift study, and the crystals on the delamanid dispersions were not identified, so the step from gastric crystals to lower release is inferred [elkhabaz2019a X8; nguyen2023c X11]. The 50% posaconazole dispersion also lost its supersaturation without an acid stage, so its crystal burden is not the only cause [elkhabaz2019a C44].

← Concepts and formulas

concept · as used in Crystallization tendency

crystallization during dissolution

Crystallization of an amorphous drug while it dissolves, either on the dissolving solid or from the supersaturated solution it creates, so that the solution never reaches the calculated amorphous solubility ilevbare2013c C20, C24. The induction-time runs of the chapter avoid the first route by starting from a solution ilevbare2013c C24. Seeds crystallized from a felodipine–HPMCAS melt in pH 3 buffer for up to a week are the dispersion route of the seed study [schram2016 P3, R10]. New aspect (Supersaturation from ASD dissolution): the frame of two routes (crystallization of the solid matrix and of the solution) and the claim that their kinetics relative to dissolution "in large part shape" the concentration–time profile alonzo2010 C32, C47; for nifedipine both routes pass through a metastable form first raina2014b C29. For resveratrol dispersions crystallization "appeared to occur in or at the surface of the matrix", also where an undissolved polymer blocked release, and the concentration reached depended on the extent of crystallization, the drug–polymer interactions and the amount of dispersion added wegiel2015 C16, C48. New aspect (Release mechanism and LoC): in the release papers of 2020–2022 crystallization on the dissolving compact ends release that began well. Heating 10% atazanavir–PVPVA compacts to 55 °C, near the drug's wet Tg, restored release, but drug release ceased at about 70 min at 64% released while PVPVA dissolved completely, and crystals were seen by PLM at 90 min; crystallization "is likely" why release stopped [correasoto2022b R5, C14, M1]. The TPGS dispersion crystallized after 1 h at 55 °C [correasoto2022b R6]. For indomethacin methyl ester the route was crystallization at the compact surface saboo2020 C17. The 2025 HPMCAS papers checked and excluded it as the cause of poor weak-base release: no crystals during 1 h or for 12 h after (except loratadine–LF after about 10 h), and amorphous bedaquiline residues [bapat2025a R6, C17; kaur2025 R14].

Findings

Crystallization from solution: F2, F3, F24. Supersaturation from ASD dissolution: F1, F2, F3, F16. Release mechanism and LoC: F18, F19, F32.

Related concepts

limits amorphous solubility; related to crystallization tendency and desupersaturation; part: matrix crystallization and solution crystallization; related to solution-mediated phase transformation. At the compact surface it is surface crystallization; ends congruent release; the test temperature is set against the wet glass transition temperature.

Formulas

Noyes-Whitney equation, classical nucleation theory.

How it is measured or obtained

The maximum concentration reached on dissolving a solvent-evaporated or spin-coated amorphous solid by in-situ UV, compared with the calculated value; polarized-light microscopy of melt-quenched particles wetted with buffer [ilevbare2013c P7, P8, R2, R4, R5; ilevbare2012b P5, P6]. In the ASD dissolution papers: concentration–time profiles read side by side with in situ solid-state measurements of slurries (Raman, MCR) and polarized light alonzo2010 P5; raina2014b P6, P10. In the release-mechanism papers: polarized light microscopy of the compact after the run, besides the stop of drug release while the polymer keeps dissolving [correasoto2022b R5].

Values in the library
materialvalue or classconditionspaper ids
ritonavir; efavirenz; celecoxibmaximum 19.8; 12.3; 2.5 µg/mL against calculated 20.6; 19.8; 22.6pH 6.8, 37 °Cilevbare2013c R2
amorphous celecoxib particlebirefringent spot within about 1 min, extensive crystals at 6 minbuffer, 20×ilevbare2013c R4
amorphous efavirenz particlesshrank over 3 h, no birefringencebufferilevbare2013c R5
felodipine slurry without polymer (ASD dissolution chapter)more than 50% crystalline at the first acquisition (1 min) at 37 °C; about 8 min to 50% at 25 °CpH 6.8alonzo2010 R2
danazol (cited)amorphous danazol reached less than a quarter of the expected concentrationrefs in the card[jackson2014 R27]
felodipine–HPMCAS melt (10% w/w)small rounded features, steps 0.77 nmpH 3, up to a week[schram2016 R10, R12]
indomethacin, neat amorphousconcentration maximum (≈11.2 µg/mL) near the onset of conversion to αpH 2, 37 °Calonzo2010 R13, R7
nifedipine glassglass gone by ≈6 min, γ then α; solution ≈6 µg/mLpH 6.8, 25 and 37 °Craina2014b R6, R14
nifedipine–HPMC 50:50≈100 µg/mL at ≈22 min, ≈41 µg/mL at 150 min5 mg/mL, 37 °Craina2014b R14
resveratrol–HPMCAS 25%, pH 6.8 against pH 1.2Raman ratio ≈0.96 and ≈0.93 after a day37 °Cwegiel2015 R17
neat amorphous celecoxib≈1.70 against ≈1.85 µg/mL for the crystal at 16 hpH 6.8, 37 °Cxie2015 R7
atazanavir–PVPVA 10%64% of the drug and all PVPVA released; drug release ceased at ≈70 min; crystals at 90 min55 °C[correasoto2022b R5]
atazanavir–PVPVA 10% with TPGScrystals after 1 h55 °C[correasoto2022b R6]
Pitfalls and disagreements

For efavirenz the shrinking particles suggest crystallization from the solution after dissolution, not on the solid ("suggesting") ilevbare2013c C23. The theoretical and experimental amorphous solids were made by different routes (melt-quench for the water sorption feeding the calculation, solvent evaporation for the measurement) ilevbare2013c X17. The Tg of the hydrated solid explains the felodipine temperature effect only in part; indomethacin crystallized more slowly despite a similar Tg drop alonzo2010 C34, C42. Supersaturation in the nifedipine slurries was not measured raina2014b X10. (synthesis: the higher temperature that restored release of the 10% dispersion also brought crystallization within the 90-min run) [correasoto2022b R5, R6].

← Concepts and formulas

concept · as used in Crystallization tendency

crystallization from the supercooled melt

Crystallization of a drug melt as it is cooled below Tm, or while it is held below Tm; it is the route probed by the DSC classification and the one relevant to melt extrusion and melt quenching baird2013 C1; vaneerdenbrugh2010a P10.

Findings

F9, F14.

Related concepts

contrasts with cold crystallization; measured by crystallization tendency classification; depends on degree of supercooling and melt viscosity; occurs in supercooled liquid.

Formulas

Barandiaran-Colmenero critical cooling rate.

How it is measured or obtained

DSC cooling at fixed rates baird2010 P1; rapid heat–cool DSC baird2013 P2; isothermal holds of the undercooled melt moseson2020 R7; rheometry while cooling baird2012 P1.

Values in the library
materialvalue or classconditionspaper ids
51 molecules23 crystallize on cooling at 20 °C/minDSCbaird2010 R1
vemurafenibcrystallized from 248.9 °Ccooling at 10 °C/minlu2016 R1
posaconazole, bicalutamide; indomethacincrystallization within 1 h; none in 4 hisothermal holds at 130–180 °Cmoseson2020 R7
acetaminophen with polymersrank orders for nucleation and growth inhibition differ, and differ from solutionundercooled melt (cited)trasi2015 R14, C28
Pitfalls and disagreements

Solid-state polymer rankings need not hold in solution trasi2015 C28. Melt and solvent classes agree only for rapid solidification vaneerdenbrugh2010a X7.

← Concepts and formulas

concept · as used in Crystallization tendency

crystallization inhibition

The delay or slowing of crystallization of an amorphous drug by a polymer mixed with it. Several explanations are in circulation and none is accepted: reduced molecular mobility through a higher Tg (antiplasticization), a lower driving force through a lower chemical potential, a higher barrier, and drug–polymer specific interactions that disrupt the drug's self-association [taylor1997 C3, C4, C5; marsac2006b C2, C5; kestur2010a C29; rumondor2009a C2]. For indomethacin and PVP the proposal is that a nucleus of the stable γ form is built of acid dimers, so that nucleation "would require" breaking the PVP–drug hydrogen bond taylor1997 C43. The effect is measured on different quantities: the overall crystallinity of a film over days (the amorphicity index), the growth rate of crystals into the supercooled liquid, or nucleation separately vaneerdenbrugh2010b E1; kestur2010a P4. In later designs the effect is read as the amorphicity index of 105 spin-coated pairs of 15 drugs and 7 polymers [vaneerdenbrugh2011a R3], as the highest polyphenol content that stays X-ray amorphous and the days to crystals at five storage conditions wegiel2013a R8, R9, and as the weeks to crystals of lumefantrine dispersions hiew2022a R6. The ranking of polymers follows complementary groups of drug and polymer (see F24). In the glass, melt and solution (formerly the page "crystallization inhibition by polymers", from ASD process, Crystallization from solution, Physical stability and Supersaturation from ASD dissolution): delay of nucleation and slowing of growth of a drug by a polymer in the glass, melt or solution. Two settings matter here. In solution, dissolved PVPVA prevented nucleation of indomethacin for more than 12 h and HPMCAS prevented that of bicalutamide for 8 h [moseson2020a C11; moseson2021 R3]. In the solid or melt, a polymer can stabilise the amorphous drug against recrystallisation and let degradation be tested on a dispersion moseson2020 C32, C55. A polymer inhibits in the solid only when intimately mixed with the drug in one phase, usually inferred from a single Tg [rumondor2009c C26; marsac2010 C31]. The polymer's help in solution does not carry into the hydrating solid or across drugs: HPMCAS slowed seeded growth of posaconazole in FaSSIF [shah2024 R22], but in the hydrating solid cryomilled and high-loading bicalutamide–PVPVA dispersions still crystallized moseson2022b R4, R6; and coated erlotinib–HPMCAS held 48 weeks at 43% RH where coated erlotinib–PVPVA failed at 6 weeks, a difference the authors attribute to the polymer moseson2022a R11, R12, C31. New aspect (Crystallization from solution): the delay of crystallization from a supersaturated aqueous drug solution by a dissolved polymer, through slowing nucleation, growth or both; for felodipine and HPMC both were slowed at µg/mL levels and nucleation far more [alonzo2012 C46]. The papers separate it into crystal growth inhibition and nucleation inhibition because polymers rank differently in the two (F6) ilevbare2013c C40. The solid-state version, in amorphous dispersions, is described at the start of this definition. New aspect (Physical stability): in dry felodipine films a few percent of polymer lowered nucleation by orders of magnitude, independent of polymer type and of any Tg rise, and the authors speculate that it raises the kinetic barrier to nucleation konno2006 C13, C14, C40. With water present the polymer still inhibits, competing with the water it brings, but the polymers then differ konno2008 C13, C10. New aspect (Supersaturation from ASD dissolution): one polymer can act differently on the two routes by which an amorphous solid loses supersaturation: PVP delayed crystallization of the wetted felodipine solid at least as well as HPMC and HPMC-AS but held felodipine solutions poorly alonzo2010 C38; the authors therefore "speculate" that some drugs "might" be formulated better with two polymers, one per route alonzo2010 C46. Because 1 mg/mL of each polymer left the crystalline solubility of felodipine unchanged, the higher concentrations from the dispersions are read as kinetically maintained supersaturation, not solubilization [konno2008a R1, C17, C22]. The papers of 2015–2018 add that a fast-releasing polymer can be paired with a minor inhibitor that then works in solution [xie2015 C6; xie2016a C69], that against matrix crystallization the inhibitor must stay in contact with the drug in the particle [xie2016a C54], and that the polymer content needed to stop matrix crystallization depends on the polymer wegiel2015 C11.

Findings

Miscibility and drug-polymer interactions: F1, F9, F10, F11, F12, F13, F15, F16, F17, F23, F24, F25, F26. ASD process: F25, F27, F31, F38, F41, F44. Crystallization from solution: F6, F7, F8, F13, F14, F15, F16, F20, F25, F26, F27, F36, F37, F41, F43, F44. Physical stability: F1, F3, F4, F10, F11, F15, F17, F18, F19, F21, F22, F24, F26, F27, F28, F29, F33, F35. Supersaturation from ASD dissolution: F2, F3, F4, F5, F6, F12, F16, F20.

Related concepts

includes crystal growth inhibition and nucleation inhibition; depends on drug-polymer interactions, drug-polymer miscibility, antiplasticization and thermodynamic driving force for crystallization; contrasts with crystallization tendency of the neat drug; limited by matrix crystallization; related to nucleation, crystal growth rate, kinetic stabilization, polymer hydrophobicity and supersaturation. Acts through kinetic barrier to nucleation and polymer rejection at the crystal growth front (proposed); not through antiplasticization at low content; lost by moisture-induced phase separation; decided by hydrogen-bond donor and acceptor groups and acid-base interaction; related to drug-polymer complexation.

Formulas

amorphicity index, viscosity-limited normal growth law, crystal growth inhibition effectiveness ratio, relative nucleation rate from seeded and unseeded induction times, induction time decomposition, empirical log-linear growth-rate equation, classical nucleation theory, supersaturation ratio, overall mass crystal growth rate.

How it is measured or obtained

Spin-coated films of drug and polymer at several ratios, scored by polarized light microscopy at day 0, 1, 3 and 7 and expressed as an amorphicity index vaneerdenbrugh2010b P1, P2, E1; hot-stage growth of spherulites in melt-quenched films with low polymer levels after nucleation at 40 °C kestur2010a P2, P3, P4; rotary-evaporated bulk powders by DSC crystallinity vaneerdenbrugh2010b P8, E5. Only growth, not nucleation, was measured in the felodipine and bifonazole–nimesulide studies kestur2010a X10; kestur2011 P3. Induction time and seeded desupersaturation in solution; DSC of undercooled melts for the dispersion (a melting peak on reheating means nucleation during the hold); matrix crystallisation by microscopy and Raman of particles in buffer [moseson2021 P12, P15, P16; moseson2020 P7]. In the crystallization-from-solution papers: seeded growth rates and unseeded induction times with and without predissolved polymer under identical conditions, combined into a relative nucleation rate (F5, F6) [alonzo2012 P10, E9, E10]; induction times by UV extinction or absorbance ilevbare2013c P9; pH-shift precipitation followed by polarized light, WAXS and HPLC hsieh2014 P2–P7; fluorescence and SHG for droplet and crystal read-outs [jackson2014 P7, P10]. In the physical-stability papers: nucleation-site counts in films konno2006 P2; marsac2006a P4; konno2008 P4, melt or film growth rates kestur2010b P3; rumondor2010 P5, PXRD crystallinity of stored powders [rumondor2009b P5]. In the ASD dissolution papers: non-sink dissolution of dispersions or of amorphous drug with polymer in the medium; solvent-shift solutions with predissolved polymer; induction times by extinction or pyrene; Raman of slurries for the solid [konno2008a P3, P5; alonzo2010 P2, P4, P5; jackson2015 P3; ilevbare2013d P6].

Values in the library
materialvalue or classconditionspaper ids
8 drugs, 7 polymersmean amorphicity index PVP = PVPVA 70 > E100 67 > HPMC 66 > HPMCAS 62 > PAA 49 > PSSA 48spin-coated films, 7 days dryvaneerdenbrugh2010b R5
amides, bases against acids with PAA and PSSAhighest for amides (64–78) and bases; lowest for acids (8–31)as abovevaneerdenbrugh2010b R3, C3
felodipine, 3% PVP, PVP/VA, HPMCAS, PVAcgrowth ratio at 70 °C 10.0, 6.5, 4.4, 1.2hot stagekestur2010a R6
bifonazole with 5% PAA5-fold at 100 °C, 15-fold at 18 °Chot stagekestur2011 R7
nimesulide with 5% PVP, PVP/VA, PVAcabout 21-, 16-, 3-fold at 18 °Chot stagekestur2011 R9, X7
50/50 powders, acidic drugs with PAA against PVPabout 64–117% against at most about 3% crystallinerotary evaporation, DSCvaneerdenbrugh2010b R9
acids, acidic N–H drugs, alcohols (13 drugs)AI with E100, PVP, PVPVA about 54–100; with PAA, PSSA mostly about 4–31spin-coated films, 7 days dry[vaneerdenbrugh2011a R3, C20, C25]
naringenin, quercetinhighest X-ray amorphous content: E100 about 85 and 70%, PAA about 30 and 20%rotary evaporation, PXRDwegiel2013a R8
naringenin, quercetin with E100 and PVPno crystals for >200 to >450 days at five conditionsPXRDwegiel2013a R9
lumefantrine–PVPVA; –enteric polymerscrystals at 1–8 weeks; only at 50% drug loading (1–40 weeks)40 °C/75% RHhiew2022a R6
IDM, PVPVA in solutionno nucleation for more than 12 h at 5–50 µg/mL; about 40 min withoutS = 6[moseson2020a R3, C11]
BCL, PVPVA 70 µg/mL and HPMCAS 1 mg/mL in solutioninduction above 1 h; none in 8 h; without polymer under 30 min37 °C[moseson2021 R3]
BCL–PVPVA dispersion in the slurryHPMCAS in the medium delays crystallisation by about 10 min and leaves form 2 growth unchangedRaman[moseson2021 R19, R20, C51]
neat PCZ and BCL meltsnucleate within 1 h between Tg and Tm; IDM did not in 4 hDSC holdsmoseson2020 R7
twenty commercial polymers on ritonavir growth (outside)five effective: Pn-IPAAmd > HPMCAS > HPMC > CAPh > CMCABseeded growth[ilevbare2012c C22]
bifonazole in melt-quenched films (outside)PAA the most effective polymer; PAA ineffective on acetaminophen in solution5 wt%kestur2011 C9; trasi2015 C26
erlotinib–HPMCAS against erlotinib–PVPVA, coated, 43% RHnone at 48 weeks against about 6 weeks40 °C, SEMmoseson2022a R11, R12
felodipine, 1 µg/mL HPMCgrowth ratio 0.607; induction time 15.3 → 136.5 min; J/J_0 7.05 × 10⁻⁴S 10, 25 °C[alonzo2012 R5, R16, R17]
ritonavir, CAB Adp 0.81induction time ≈120 → ≈244 min5 µg/mL, 20 µg/mL drugilevbare2013c R9
celecoxib, HPMC2.4 → 12.6 min (22 µg/mL); 27.5 → 855.5 min (11 µg/mL)5 µg/mL, 37 °Cilevbare2013c R9, R15
danazol, HPMC-AS6–10 → 250–516 min10 µg/mL polymer, 25 °C[jackson2014 R17]
warfarin, six polymersamorphous precipitate ≥ 6 h; 103–224 µg/mL1 mg/mL, pH shifthsieh2014 R4
celecoxib, 5 µg/mL HPMCASsupersaturation held 16 h; J_0/J_p about 4,00022 µg/mL drug[xie2016a R3, R13]
six dihydropyridines, 4.5 mg/mL polymerPVPVA = HPMC = HPMCAS > PVP > CMCAB = CAPAdp > P2VP = PAAnominal 10 mg/mLraina2015b C16, R9
telaprevir 150 µg/mL, 5 µg/mL polymerHPMC 243, HPMCAS 175–226, CA-Sub-090 106 min against 14–15 withoutpH 6.8, 37 °C[mosqueragiraldo2016 R4, R5]
acetaminophen S 1.5, 10 µg/mL95% of vials: none 26, PAA 11, HPMCAS 470, HPMC 1043 min20 °Ctrasi2015 R1
atazanavir, volunteer media V5 and V6, 1 mg/mL HPMC134 ± 23 and 45 ± 7 → >360 minpH 6.5, 37 °C[enright2020 R28]
telaprevir δ 28.8, 150 µg/mL HPMCAS-MF, four lecithin mediaabout 357–474 min, no difference37 °C[lu2018 R7]
enzalutamide 120 µg/mL, 50–100 µg/mL HPMCASabout 431–530 min; P177 about 71–125 minpH 6.5wilson2021 R3
felodipine with 3% PVP, HPMCAS or HPMCnucleation ≈1.3 × 10⁵ → ≈2–3 × 10³ m⁻³ s⁻¹, equal for all three0% RH, 22 °Ckonno2006 R4, C14
nifedipine and felodipine with 7 wt% PVPabout two orders of magnitude lower; 3–4-fold more to 25 wt%0% RHmarsac2006a R16, C18
felodipine with 3% PVP vs HPMCAS≈8.6 × 10⁵ vs ≈1.3 × 10⁴ m⁻³ s⁻¹75% RHkonno2008 R3.b, C10
felodipine with 3 wt% PVP K-90growth 21-fold slower at 70 °C, 6-fold at 110 °Cmeltkestur2010b R8
felodipine with 25–50% PVP vs HPMCAS51–82% vs 9–28% crystallized after ≈490 days75–93% RH[rumondor2009b R9, C13]
acetaminophen with PAA or HPMCASPAA the best growth inhibitor but most nuclei; HPMCAS the reverse10% w/wtrasi2012a C46, C53
flutamide with acceptor or donor polymersacceptor polymers (PVP, PDMA, PIPA) slow growth; donor polymers do not10 mol%trasi2012b C8, C50
felodipine forms I and II with 3% PVPrate ratios ≈15.8 and ≈18.7 at 60 °C, ≈3.5 and ≈6.5 at 100 °Cmelt growthkestur2013a R15
celecoxib with PVP and TPGSslower than PVP alone at 70–120 °C at 10% PVPmelt growthmosqueragiraldo2014 R2.b, C13
resveratrol and curcumin dispersionsPVP and E100 best for resveratrol; E100 best, PVP worst for curcuminstorage at 84% RHwegiel2013b R12; wegiel2014 R13
ATP(S) vs ATP(R) with 5% HPMCAS-HF or -LFS 1.9–2.5 times faster than R (HF); R 1.5–1.9 times faster than S (LF)melt growth, 50–80 °C[sato2015 R5.i–l]
six compounds with 5% PHBratio with/without polymer ≈0.42–10, rising with compound Tgmelt growthsato2016 R11
indomethacin α vs γ with 10% HPMCratio without/with polymer ≈2900 vs ≈2.9 at 120 °Cmelt growthtian2017 R12.i
tacrolimus–HPMC 1:1 vs pure amorphous drugamorphous at 4 weeks vs crystalline after 2 weeks at 40 °C/75% RHopen dishtrasi2017 R16, C40
felodipine dispersions 75% HPMCAS / HPMC / PVPmaxima 14.1 / 4.9 / 2.3 µg/mLpH 6.8, 37 °C[konno2008a R7]
felodipine 9.5 µg/mL, 150 µg/mL polymer8.5–9.1 (HPMCAS), 7.8–8.9 (HPMC) µg/mL held; PVP to ≈2.8solvent shift[konno2008a R14]
amorphous indomethacin, 250 µg/mL PVP≈23.8 µg/mL held 4 h at 25 °CpH 2alonzo2010 R14
indomethacin solvent shift, 37 °Cnone ≈42 → ≈7.3 µg/mL in 30 min; HPMC ≈39 held about 2.5 hpH 2alonzo2010 R17
danazol dispersions PVP / HPMC / HPMCASinduction ≈56–70 / ≈188–200 / ≈284–319 minpH 6.8, 25 °C[jackson2015 R5]
ritonavir 20 µg/mL, 5 µg/mL of 15 additivesinduction 23.6 to 242.8 against 119.3 minpH 6.8, 37 °C[ilevbare2013d R12]
nifedipine γ → α, 1 mg/mL polymerγ intact ≈20 (PVP), ≈60 (HPMC), ≈110 min (HPMCAS) against ≈1325 °Craina2014b R9
celecoxib 22 µg/mL, 5 µg/mL PVP-K12 / PVP-K29/32 / HPMCAS, HPMC / PAAonset ≈1 h / ≈2 h / more than 8–16 h / as no polymerpH 6.8, 37 °C[xie2015 R1; xie2016a R3]
tacrolimus 50 µg/mL with 50 µg/mL HPMConset 6–8 h against within 3 hpH 4.5, 37 °Ctrasi2016 R9
indomethacin, 100 µg/mL HPMC against EPOheld ≈1.5 h against crystals during the titrationpH 2.2, 37 °C[xie2017 R5, R8]
resveratrol dispersionsno matrix crystallization from 80% HPMC, 85% HPMCAS, 95% CMCABpH 6.8, 37 °Cwegiel2015 R14

Evidence table: crystallization inhibition.

Pitfalls and disagreements

Inhibition of growth, of nucleation and of overall crystallinity need not coincide: the best growth inhibitor of acetaminophen promoted nucleation trasi2012a C46. Polymers rank differently for different drugs vaneerdenbrugh2010b C20; kestur2011 C29. No polymer property tested at low levels (Tg, melting point, molecular weight) explained the order kestur2010a C32, C34, C35. The hydrogen-bond mechanism is a rank correlation over four polymers, and the demixing step is the authors' "speculation" kestur2010a C38, C42. Film results carried over to bulk powders for six of seven drugs vaneerdenbrugh2010b C42, C43, C44. The IR ranking of the polyphenol polymers separated the best and worst but not the middle ones, and polymer content differed from 30 to 80% in the stability comparison wegiel2013a X4, X3. The same ionic contact that protected lumefantrine dispersions slowed the dissolution of the enteric polymers hiew2022a C53. A polymer that inhibits in solution does not necessarily inhibit in the hydrating solid (HPMCAS and bicalutamide; polymers predissolved with amorphous nifedipine) [moseson2021 C51; raina2014b C11]. The mixing condition fails when the dispersion has phase separated, so water-induced separation can remove the benefit of the polymer [rumondor2009c C6]. Celecoxib with HPMC or HPMCAS at 22 µg/mL and 5 µg/mL polymer gave induction times of about 10–13 min in one laboratory and more than 8–16 h in another [ilevbare2013c R9; xie2015 R1; xie2016a R3]. Growth and nucleation data for the same polymers are often measured at different supersaturations ilevbare2013b X15. A polymer that raises the crystalline solubility or lowers the onset of phase separation changes the supersaturation it is credited with (F19) [jackson2014 X9]. In the solid state, equal effectiveness holds for felodipine nucleation when dry, not for growth (polymers differed several-fold) or in humid air konno2006 C14; kestur2010a R6; konno2008 C10. After moisture-induced phase separation the PVP level stopped mattering for crystallization at 93% RH [rumondor2009b C20]. The mechanism (barrier at the nucleus, hydrogen-bond coupling, polymer encaged in the crystal) was argued, not measured konno2006 C40; kestur2010b C35, C42. A polymer's effects on nucleation and on growth need not agree trasi2012a C53. Equal inhibition of two polymorphs is read as action in the liquid, unlike adsorption in solution kestur2013a C6, C28. PVP slowed felodipine bulk growth about sevenfold but surface growth only twofold kestur2013b C32. A polymer can also speed growth (PHB with high-Tg compounds) sato2016 C29; the factor depends on the polymorph and on the side of the growth maximum tian2017 C37, C35; and two grades of one polymer can favour opposite enantiomers [sato2015 C28]. PAA, the most effective polymer for bifonazole in melt-quenched films and the best growth inhibitor for acetaminophen melts, was ineffective on acetaminophen in solution (synthesis: contrast between papers) kestur2011 C9; trasi2012a C46; trasi2015 C26. In the ASD dissolution papers the ranking depends on the drug (PVP worked for indomethacin, not for felodipine, danazol or ritonavir) [alonzo2010 C22; konno2008a R7; jackson2015 R5; ilevbare2013d R12]. Solution screens predict only the rank order, and HPMC was misranked for felodipine [konno2008a C31; jackson2015 X8]. An additive can stabilize droplet size yet promote crystallization [ilevbare2013d R14].

← Concepts and formulas

concept · as used in Crystallization tendency

crystallization tendency classification

A sorting of neat drugs by when, if ever, they crystallize under a fixed protocol. From the melt: class I crystallizes on cooling (split into I-A, which still crystallizes on liquid-N2 quenching, and I-B, which vitrifies), class II only on reheating, class III on neither baird2010 R1, C10; the labels very high, high, moderate and low tendency are used for I-A, I-B, II and III baird2013 P5. After rapid solvent evaporation: rapid, intermediate and slow crystallizers vaneerdenbrugh2010a P9. An extended scheme combines nucleation behaviour and growth rate into classes IA, IB, II, III and IV trasi2014a C17.

Findings

Crystallization tendency: F5, F9, F10, F14, F17, F18. Physical stability: F17, F24.

Related concepts

measures crystallization tendency; extended by extended nucleation and growth classification; related to glass-forming ability, glass stability, critical cooling rate and nucleation and crystal growth.

Formulas

none specific; the critical-rate formulas refine the classes: Barandiaran-Colmenero critical cooling rate.

How it is measured or obtained

DSC heat–cool–heat at 20 and 10 °C/min plus liquid-N2 quench baird2010 P1, P6; spin coating and polarized light microscopy over 7 days vaneerdenbrugh2010a P8, P9; growth rates for the extended scheme trasi2014a P1.

Values in the library
materialvalue or classconditionspaper ids
51 molecules23 class I, 11 class II, 17 class IIImelt DSCbaird2010 R1
93 drug–solvent combinations39 class I, 23 class II, 31 class IIIspin coatingvaneerdenbrugh2010a R9
felodipineclass III (melt and solvent)both routesbaird2010 M37; vaneerdenbrugh2010a R8.ag
nifedipineclass IImelt DSCbaird2010 M32; raina2015b C12
celecoxibclass II (melt); class III (solvent)both routesbaird2010 M26; vaneerdenbrugh2010a R8.ae
atenololclass I (melt); class III (solvent)both routesvaneerdenbrugh2010a R17
cilnidipine, nisoldipine, nitrendipineclass IIImelt DSCraina2015b C12
curcumin; resveratrolintermediate; rapid crystallizersolvent schemewegiel2014 R21; wegiel2013b C6
naringenin; quercetinintermediate; rapidsolvent schemewegiel2013a C19
lumefantrineclass 3melt DSCtrasi2020 R5
pretomanid, delamanidclass Imelt DSCnguyen2023a C59
nimesulide, nilutamide, celecoxibextended class II (fast growth, low nucleation)extended schemetrasi2014a C17
celecoxibslow crystallizer by the cited scheme, yet growth "quite fast"; "quite hard to nucleate"melt filmsmosqueragiraldo2014 R17, R16, R15, C7
acetaminophenforms a glass because nucleation and growth zones barely overlapDSC and hot stagetrasi2012a C39
Pitfalls and disagreements

The class can be altered by the heating and cooling rates trasi2012a C7, and 7 of 28 class II and III molecules changed behaviour at other rates baird2010 X12. Two schemes are mixed in some papers trasi2014a X10. Celecoxib is called a "slow crystallizer" by one later paper although the 2010 screen placed it in class II mosqueragiraldo2014 R17; baird2010 M26. The melt class predicts crystallization from aqueous solution only partly raina2015b C28. A slow class can hide a fast grower: celecoxib is class-slow because it nucleates poorly, not because it grows slowly mosqueragiraldo2014 C7.

← Concepts and formulas

concept · as used in Crystallization tendency

crystallization tendency

The propensity of an amorphous or supercooled drug to crystallize, expressed as a class from the melt or after solvent evaporation, as a growth rate, or as a time to crystallize in storage or solution baird2010 C8; vaneerdenbrugh2010a C33; trasi2014a R1. The inherent crystallization tendency is called the most important parameter of physical instability and is said to depend on conformational flexibility, configurational entropy, driving force, viscosity and hydrogen bonding, factors "not fully elucidated" trasi2017 C6. New aspect (PEG dispersions): the tendency can also be read in a dispersion, relative to the neat drug; in 20/80 comelts with PEG 3350 at 25 °C PEG delayed benzocaine, left haloperidol unchanged and brought forward ibuprofen and fenofibrate, so the authors conclude that PEG "can delay, promote or have no influence" on drug crystallization and that both the neat-drug tendency and the interaction with PEG matter zhu2012 C6, C38, C39. New aspect (solution methods): in solution work the tendency decides whether a supersaturated drug first phase-separates or first crystallizes; very fast crystallizers (nifedipine; nimodipine at 37 °C) crystallize just below the LLPS concentration unless a polymer slows nucleation, slow crystallizers form a drug-rich phase first raina2015a R17, C11, C37; ilevbare2013a C27. New aspect (Miscibility and drug-polymer interactions): the tendency of the neat drug carries into polymer dispersions. In a spin-coating screen of eight drugs with seven polymers, drugs whose pure amorphous form crystallizes rapidly (class I) "tended to be" more difficult to stabilize and, the authors say, needed more polymer; the overall crystallization tendency of a drug in the dispersions is related to that of the pure amorphous drug prepared in a similar way vaneerdenbrugh2010b C1, C15. Class I compounds are therefore "ideal model compounds" for telling polymers apart, whereas the compounds usually used as models in solid-dispersion work are often class III vaneerdenbrugh2010b C16, C17. Knowledge of the pure drug's tendency is called "extremely important" early in development and a guide to whether a dispersion is feasible vaneerdenbrugh2010b C18. Further cases: naringenin (intermediate crystallizer) and quercetin (rapid crystallizer) could not be made amorphous by rotary evaporation, naringenin only as a spin-coated film, and needed 15–80% polymer to give an amorphous dispersion wegiel2013a C19, R7, R8; the 15 model drugs of the donor–acceptor screen all have an intermediate to rapid tendency [vaneerdenbrugh2011a C16]; lumefantrine is a class III glass former (no crystallization on heat–cool–heat; Tg 19.7 °C) whose PVPVA dispersions crystallized in 1–8 weeks hiew2022a R3, R6. New aspect (ASD process): in process papers the tendency is a precondition: a drug must resist crystallization while the solvent leaves in granulation, so slowly crystallizing, larger and more complex molecules are "most suitable" for solvent granulation trasi2019 C35; ritonavir crystallizes slowly, so amorphous phase separation can be studied without crystallization interfering [li2020 C15]. Bicalutamide is "a fast-crystallizing compound in solution" and also crystallizes inside the hydrating solid; the authors name its low Tg, high crystallization tendency, high loading and fast polymer dissolution as the reasons for its susceptibility to matrix crystallization [moseson2021 C6, C60]. The tendency decides whether a residual-crystal level is tolerable: posaconazole, a fast crystallizer, lost supersaturation with 0.3% crystals, whereas indomethacin–PVPVA, a slow crystallizer, was little affected (cited) [shah2024 C56, R26]. Spray-dried ezetimibe–HPMCAS crystallized within 12–19 days at 40 °C/75% RH uncoated duong2022c R16, R17. New aspect (Bioperformance): the tendency is used as a descriptor to explain why felodipine reaches LLPS and nifedipine crystallizes first raina2014a C31, R20. Pretomanid is a low-Tg, class I rapid crystallizer (reduced glass transition temperature 0.67): its melt-quenched glass crystallized on heating just above Tg and within minutes at room temperature, and its solution crystallized within 2 min near the amorphous solubility nguyen2023a C10, C59, C61, R1, R3, R4. New aspect (Crystallization from solution): the solution counterpart of the class from the melt, that is how fast a drug crystallizes from a supersaturated aqueous solution, read as the type 1 or type 2 behaviour of a pH shift, as the precipitate form (crystalline against amorphous), or as an induction time at a defined supersaturation hsieh2012 C42; ilevbare2013c R8; hsieh2014 R4. It is only partly predicted by the melt class (see Crystallization tendency F18). The synchrotron WAXS ranking of six dihydropyridine precipitates (nifedipine 0.25 h to cilnidipine 12 h) is wider than their melt classes (two Class II, four Class III) raina2015b R1, R6. New aspect (Physical stability): the relative crystallization tendency of drugs from dispersions "may be dependent" on that of the pure drugs: nifedipine stayed the faster nucleator over felodipine in PVP at every content, dry and humid marsac2006a C52; marsac2008a C7. New aspect (Supersaturation from ASD dissolution): behaviour of neat amorphous drugs on wetting: nifedipine, a "relatively fast crystallizer" (cited), crystallized in the particle within minutes even with polymer in the medium, and its faster inherent crystallization than felodipine "appears" to carry over to water raina2014b M1, C10, C28; Tg alone did not rank felodipine and indomethacin alonzo2010 C42. Resveratrol, classed as a rapid crystallizer (cited), crystallized in or at the matrix of cellulosic dispersions within the first hour of slurrying, and its high melting point and rapid crystallization make effective polymeric inhibitors "essential" wegiel2015 R22, C27, C16; lopinavir, a slow crystallizer from solution, was chosen so that leaching and solubility could be measured over hours without crystals [li2016b C5; li2018 C11].

Findings

Crystallization tendency: F9, F13, F14, F17, F18, F20. PEG dispersions: F8, F9. Solution methods: F8, F9. Miscibility and drug-polymer interactions: F15, F16, F23, F24, F26. ASD process: F22, F25, F27, F29, F31, F41, F43, F44. Bioperformance: F1, F10, F27. Crystallization from solution: F1, F3, F16, F21, F35, F36. Physical stability: F2, F17, F24, F33, F34. Supersaturation from ASD dissolution: F1, F2, F16, F23, F29.

Related concepts

measured by crystallization tendency classification; depends on nucleation, crystal growth rate, melt viscosity and thermodynamic driving force for crystallization; related to glass-forming ability, glass stability, physical stability, stochastic nucleation, nucleation induction time, heterogeneous nucleation, semicrystalline solid dispersion, liquid-liquid phase separation, crystallization and crystallization inhibition. Governs matrix crystallization; moderated by crystallization inhibition by polymers. Related to solution crystallization and reduced glass transition temperature. Measured in solution by chaser and non-chaser behaviour; related to conformational flexibility and nucleation rate; related to glass transition temperature.

Formulas

Hoffman equation, classical nucleation barrier; amorphicity index. Reduced glass transition temperature, printed in words for pretomanid (0.67; recomputed 0.673) nguyen2023a E2, M1.

How it is measured or obtained

By the DSC cooling–reheating classification baird2010 P1; by spin coating and dry storage vaneerdenbrugh2010a P9; by seeded growth rates trasi2014a P1; by induction times or precipitate crystallinity in supersaturated solution ilevbare2013c R8; raina2015b R6; by storage of neat glasses baird2010 P7; nguyen2023a R4. In dispersions: the fraction P(t) of 50 neat melts that had started to crystallize at 25 °C, followed for about 500 h zhu2012 P3; onset and t_{50} of the drug in the comelt by XRD and by two Raman bands zhu2012 P5, P6, R23; simultaneous SAXS/WAXS of the blend from 0.2 min to 24 h zhu2011 P2; crystallization of each neat melt cooled to room temperature under polarized light zhu2011 P7, R6. In solution: by the order of extinction onset and visible crystals, with and without a nucleation inhibitor raina2015a R4, R5, and by how long a drug-rich phase persists before the probe signal returns to the aqueous value (lopinavir 94 min; ritonavir not within 213 min) [tres2018 R20, R21]. For drug–polymer films of the spin-coating screen, by scoring crystalline area over 7 days and expressing it as an amorphicity index, averaged over drug/polymer ratios, polymers or drugs vaneerdenbrugh2010b P2, P3, E1, E2, E3, E4. The class of the pure drug is cited from the group's melt and spin-coating classifications vaneerdenbrugh2010b R11. In the process papers: isothermal DSC holds of the undercooled melt between Tg and Tm followed by reheating (a melting peak means nucleation during the hold): posaconazole 1 h at 130–160 °C and bicalutamide 1 h at 130–180 °C nucleated, indomethacin did not in 4 h at 130–150 °C moseson2020 P7, R7; in solution by induction time and seeded growth [moseson2021 P12, R3]. In raina2014a the tendency is cited, not measured: felodipine a slow and nifedipine a faster crystallizer raina2014a R20. The crystallization-from-solution papers cite the classes from cooling the melt (Baird 2010) and from rapid solvent evaporation (Van Eerdenbrugh 2010), and measure the solution tendency by the pH-shift titration or the unseeded induction time hsieh2012 P7; ilevbare2013c P9. In the physical-stability papers, by nucleation in films and crystallization on heating of melt-quenched glasses marsac2006a P4, P10. In the ASD dissolution papers: by time to crystallize in aqueous slurry (Raman) and by microscopy of wetted particles alonzo2010 P5; raina2014b P4, P6.

Values in the library
materialvalue or classconditionspaper ids
31 drugsMCV 216 to 0.02 µm/smelt growthtrasi2014a R1
haloperidol, benzocainecrystallized within seconds after quench coolingpure meltszhu2012 R1
ibuprofen; fenofibratecrystallization probability 0.20 and 0.34 after 3 weekspure melts at 25 °Czhu2012 R2, R3
ketoconazoleamorphous after 24 h at room temperaturepure meltzhu2011 R6
nifedipine against felodipinegreater tendency for nifedipinemelt-quenched, heated at 2 °C/minmarsac2006a R5, C11
propranolol against nicardipine saltspropranolol salts more prone to crystallizeamorphous saltstowler2008 C32
dihydropyridinescilnidipine slowest, nifedipine fastest in solution; for six drugs the ranking is cilnidipine < felodipine = nisoldipine < nitrendipine = nimodipine < nifedipinesupersaturated buffer; synchrotronraina2015b C13, C28
nifedipine; nimodipinenifedipine a very fast crystallizer; nimodipine faster than the other four dihydropyridinesaqueous media (cited)raina2015a R17
felodipinerelatively slow; nifedipine fastercitedraina2014a R20, C31
ritonavir, efavirenz (class III); celecoxib (class II)induction times about 120, 5.1 and 2.4 minsupersaturated buffer, 37 °C, calculated amorphous solubilityilevbare2013c M1–M3, R8
ritonavirlow tendency; drug-rich phase persists for 213 minno polymer, 24 °C[tres2018 P5, M1, C29, R20]
lopinavirdrug-rich phase lost after an induction period of 94 minno polymer, 24 °C[tres2018 P5, R21]
papaverine, warfarin, glyburide (slow); dipyridamole (rapid)slow from the melt but crystalline on rapid supersaturation; all crystallized without polymer within about 15–30 minpH shift, 25 °Chsieh2014 R1, R18, C30
pretomanidneat glass crystallized within minutesroom temperaturenguyen2023a R4, C10
drugs with polymerspure-drug tendency related to that in dispersionsspin coatingvaneerdenbrugh2010b C1
haloperidol in PEG 3350onset −100 s, t_{50} 7 s; crystalline from the first SAXS/WAXS frame20/80, 25 °Czhu2012 R23.a, R23.b; zhu2011 R9
benzocaine in PEG 3350onset 120–180 min, t_{50} 110–170 min20/80, 25 °Czhu2012 R23.a, R23.b
ibuprofen in PEG 3350onset about 100 min, t_{50} about 70 min20/80, 25 °Czhu2012 R23.a, R23.b
fenofibrate in PEG 3350onset 120–180 min, t_{50} 140–200 min20/80, 25 °Czhu2012 R23.a, R23.b
chlorpropamide; nilutamide in PEG 3350partially crystalline at 24 h20/80, 25 °Czhu2011 R18.c, R18.e
ketoconazole in PEG 3350amorphous at 24 h20/80, 25 °Czhu2011 R18.f
phenacetin, chlorzoxazone, benzamide, lidocaine, flufenamic acid (class I)mean AI over 7 polymers 47, 47, 48, 55, 59%spin-coated films, 7 days dryvaneerdenbrugh2010b R4
flurbiprofen (class I/II); chlorpropamide, bifonazole (class II)mean AI 68, 73, 97%as abovevaneerdenbrugh2010b R4
class I films with every polymer at 75/25 drug/polymeralmost all fully crystalline (AI 0)as abovevaneerdenbrugh2010b R2
naringenin; quercetinintermediate; rapid crystallizerneat films, rotary evaporation and spin coatingwegiel2013a C19, R7
lumefantrineclass III; melting onset 127.9 °C, Tg 19.7 °CDSC heat–cool–heathiew2022a R3
bicalutamide in pH 6.8 buffernucleation in under 30 min without polymer, under 5 min at the highest supersaturation; seeded growth near zero at 30 µg/mL, extremely rapid at 150 µg/mL37 °C[moseson2021 R3, R4]
posaconazole and bicalutamide neat meltsmelting endotherm on reheating after 1 h holds below TmDSCmoseson2020 R7
indomethacin neat meltonly Tg after 4 h at 130–150 °CDSCmoseson2020 R7
ritonavir and lopinaviramorphous granules unchanged for one month at 40 °C/75% RHXRPDtrasi2019 R2
uncoated spray-dried ezetimibe–HPMCAS 50 and 70% DLcrystals by SEM at 19 and 12 daysopen dish, 40 °C/75% RHduong2022c R16, R17
uncoated ASD powders of four drug–polymer systemscrystals within about 1–2 weeks40 °C, 31–75% RHmoseson2022a R11
PCZ 100 µg/mL in FaSSIF V2, seeds 10 µg/mL, HPMCAS 0 / 3 / 9 times the drug% left at 30 min about 4.5 / 10.5 / 43.5seeded solution[shah2024 R22]
nevirapineinduction about 5 min at S 3.9 (fast without polymer)pH 6.5[hate2017 R2]
atazanavirrelatively slow; crystallization inhibited more than 4 h with enough polymercited[hate2018 C13]
pretomanidinduction time under 2 min at 160 µg/mL in seven media; melting onset of the crystals 147 °CDSC, 37 °Cnguyen2023a R1, R3, R4
carvedilol, clotrimazole, clozapine, ketoconazoleclass III (melt) and III (solvent evaporation); type 2pH shifthsieh2012 M2, M3, M4, M7, R1
loratadineIII and II/III; type 2samehsieh2012 M8, R1
bifonazole; dipyridamole; droperidol; papaverine; pyrimethamineII/II; I/III; II/III; III/II; not classed; all type 1samehsieh2012 M1, M5, M6, M9, M10, R1
six dihydropyridinesmelt: nifedipine, nimodipine Class II; others Class III; solution: 0.25–12 hDSC; solvent shiftraina2015b R1, R6
nine drugs (griseofulvin, celecoxib, ezetimibe, danazol fast; telaprevir, nifedipine, nevirapine intermediate; ritonavir, atazanavir slow)induction time without polymer about 1 to 66 min100 mM phosphate pH 6.8, 37 °C; basis of the classes not given[mosqueragiraldo2018a R1, M1–M9, X11]
nifedipine vs felodipine glass0.39 vs 0.17 × 10¹⁰ m⁻³ day⁻¹; only nifedipine crystallized on heating (87 °C)22 °C; 2 °C/minmarsac2006a R4.a, R5
nifedipine vs felodipine with 25% PVP≈4.3 × 10⁵ vs ≈2.3 × 10³ m⁻³ s⁻¹75% RHmarsac2008a R1.f, R1.l
resveratrolrapid crystallizer; a polymer is needed to make it amorphousspin coating, rotary evaporation, cryomillingwegiel2013b R1, C6, C27
curcuminintermediate crystallizer, relatively high tendencysolvent evaporationwegiel2014 C9, C1
celecoxibgrows fast but nucleates poorlymelt filmsmosqueragiraldo2014 C7
tacrolimus in a marketed generic capsulecrystallized within days at 40 °C/75% RH, faster than the pure amorphous drugopen dishtrasi2017 R12, R16, C28
lumefantrine; artemetherclass 3; class 1DSC heat–cool–heattrasi2020 R5, R6, C7
felodipine, neat amorphous≈54% crystalline at 1 minpH 6.8 slurry, 37 °Calonzo2010 R2
indomethacin, neat amorphousamorphous for at least 4 hpH 2 slurry, 25 °Calonzo2010 R11
nifedipine, neat glasscrystalline (γ) by ≈6 min with or without dissolved polymerpH 6.8, 25 °Craina2014b R6, R2
resveratrolrapid crystallizer (cited); Tm 262 °Cstorage and solvent-evaporation classwegiel2015 R22, M1
celecoxib, 22 µg/mL without polymercrystallization within 5 minpH 6.8, 37 °C[xie2016a C13]
lopinavirslow crystallization from aqueous solutionpH 3[li2016b C5]
delamanidinduction time 2.4 min at pH 1.6 and 10.7 min at pH 6.5 at the amorphous solubility; no glass on cooling the melt at 50 °C/min; Tg 43.3 °C37 °Cduong2022a R2, R3
praziquantel, racemate and R-enantiomerabout 5.5 and 5 min without polymer at their amorphous solubilities; 83 min and 37 min with HPMCAS-MFPBS pH 6.5, 37 °C[polyzois2024 R3, R4]
paclitaxelunseeded crystallization within 6 min at S = 88 without polymerpH 6.8, 37 °C[que2018 R3]
Pitfalls and disagreements

The tendency depends on route and medium: rapid and slow drying differ strongly vaneerdenbrugh2010a C26, and the solid-state ranking need not hold in dilute aqueous solution hsieh2014 C30; in the dihydropyridine series, by contrast, the solution ranking and the two drugs that recrystallize from the melt coincide raina2015b C28. No single descriptor captures it vaneerdenbrugh2010a C45. In dispersions, heterogeneous nucleation by the carrier can break the link to the pure drug zhu2012 C41: the link "was found to break down for the slowly crystallizing compounds" ibuprofen and fenofibrate, because crystalline PEG apparently promotes their heterogeneous nucleation zhu2012 C41. The neat ibuprofen and fenofibrate statistics stop at 3 weeks and the dispersion runs have unstated replicates zhu2012 X5, X6. The neat-melt entries of the 2011 PEG paper (chlorpropamide, nilutamide and ibuprofen "within a few hours") come from single melts and sit beside the 50-sample statistics of the 2012 paper for ibuprofen zhu2011 R6; zhu2012 R2. The polymer fraction needed per drug is not tabulated, although the day-by-day scores show the highest drug fraction that stayed amorphous vaneerdenbrugh2010b X13, R2. The class of flurbiprofen depends on the solvent (class I from ethanol–dichloromethane, class II from ethanol), and the HPMC and HPMCAS films used the mixed solvent vaneerdenbrugh2010b X2, X12. Class III drugs were left out on the expectation of stability over a week vaneerdenbrugh2010b C13. Polymer and chemistry change the ranking within a class, so the class is a risk flag, not a prediction (see F15) vaneerdenbrugh2010b C20. The link from the neat drug to the dispersion held in the spin-coating screen vaneerdenbrugh2010b C1 but broke down for ibuprofen and fenofibrate in PEG zhu2012 C41 (synthesis: the two papers differ in carrier and test). Posaconazole, indomethacin and bicalutamide were chosen as class III glass formers, yet neat posaconazole and bicalutamide nucleated within an hour between Tg and Tm; an isothermal hold near Tm tests nucleation differently from the cooling and reheating test that defines the class (synthesis) moseson2020 C5, R7. The granulation conclusion rests on one pair of slow crystallizers trasi2019 C53. In the Bioperformance papers the tendency of the neat melt does not carry into solution with polymer (F10). A rapid crystallizer in the neat state and in solution can still be held by one polymer: HPMCAS-HF kept pretomanid in solution for over 1000 min at S = 5 while most enteric polymers failed within minutes (F27) nguyen2023a R5. In the pH-shift study class III covers all type 2 bases and also three of the four classifiable type 1 bases, so it does not separate the types hsieh2012 X1. Efavirenz is class III yet crystallizes in minutes at S about 2.3 ilevbare2013c R9.a, X4. Failing to crystallize in the DSC screen "does not guarantee" slow crystallization in solution raina2015b C29. The carry-over of the nifedipine–felodipine ranking into PVP dispersions rests on one drug pair and one polymer, and no PVP level giving equal stabilization of the two drugs was found marsac2006a X7. A favourable class did not protect a marketed product whose process left the drug outside the polymer: tacrolimus forms a stable glass, yet one generic crystallized faster than the pure amorphous drug trasi2016 C13; trasi2017 C28, C47. In the ASD dissolution papers nifedipine and felodipine have similar Tg and moisture sorption (cited) yet differ in water only when polymers are present raina2014b C27, R17; the felodipine–nifedipine comparison is cross-study raina2014b X16. Delamanid behaved like tacrolimus and bicalutamide, whose residual crystals the polymer does not poison, rather than like indomethacin duong2022a C58.

← Concepts and formulas

concept · as used in Crystallization tendency

crystallization

In this chapter, the formation of crystals from a supersaturated solution or from a drug-rich phase; it depletes the supersaturation, unlike a drug-rich phase, which keeps it [tres2017 C42].

Findings

F3, F8, F9.

Related concepts

contrasts with drug-rich phase and liquid-liquid phase separation; related to crystallization inhibition, crystallization tendency and nucleation induction time.

Formulas

none.

How it is measured or obtained

By visual particles and birefringence under polarized light raina2015a P7; by the return of a probe signal toward its aqueous value (pyrene I1/I3, PRODAN lifetime) [raina2015a R16; tres2018 R21]; by a long lifetime component of the drug itself [tres2017 R1.g]; in outside work by PXRD of compacts [bhardwaj2018 R3].

Values in the library
materialvalue or classconditionspaper ids
nifedipinecrystals in buffer at ≈38 (25 °C) and ≈58 µg/mL (37 °C)extinction onset with visible crystalsraina2015a R4, R5
nimodipinecrystals at ≈21.5 µg/mL (37 °C)sameraina2015a R4, R5
nifedipine with HPMCpyrene I1/I3 back to ≈1.60 at 400 µg/mL after ≈0.76 at 200 µg/mL25 °Craina2015a R16
felodipine 108 µM, no HPMClifetime component 9.04 ns (17%) after 15 hpH 7.4[tres2017 R1.g]
lopinavir, no polymerlifetime 1.31 ns at 94 min to 1.06 ns at 213 minpH 7.4, 24 °C[tres2018 P5, R21]
itraconazole dispersionsno crystallinity in compacts after dissolutionPXRD[bhardwaj2018 R3, C14]
Pitfalls and disagreements

Crystallization of a drug-rich phase was inferred from fluorescence in three papers and confirmed by an independent test in none of them [raina2015a X15; tres2017 X6; tres2018 X5]. Extinction cannot tell crystals from a liquid phase raina2015a C9.

← Concepts and formulas

concept · as used in Crystallization tendency

degree of supercooling

\Delta T = T_m - T, how far below its melting point a liquid is held or crystallizes; "undercooling" and "supercooling" are used as synonyms baird2013 E4; tian2017 E4. It sets the driving force, which is proportional to ΔT tian2017 C22. New aspect (PEG dispersions): in a dispersion the supercooling of each component is set by the melting point of that component in the mixture, which the other component depresses. For benzocaine the authors cite a melting point of around 89 °C for the neat drug and around 48 °C at the eutectic, so the supercooling at 25 °C is much smaller in the PEG dispersion zhu2012 C32, M3. PEG's own supercooling falls when a drug lowers its melting point zhu2011 C21, R1. New aspect (C. Amorphous solubility and LLPS): in the amorphous solubility papers the undercooling is T_m - T at the temperature of the solution experiment (25 or 37 °C), together with T_g - T_\infty, where T_\infty is the temperature at which crystal and supercooled liquid would have equal enthalpy; both are used to judge whether the Hoffman free-energy estimate is reliable almeidaesousa2015 C34, C35, R16.

Findings

Crystallization tendency: F2, F7, F10, F11. PEG dispersions: F5, F8, F10. C. Amorphous solubility and LLPS: F7. Physical stability: F18, F28, F29.

Related concepts

related to thermodynamic driving force for crystallization; used by Barandiaran-Colmenero method and critical cooling rate; part of crystallization from the supercooled melt; reduced by melting point depression; governs the accuracy of the Hoffman equation and the amorphous-crystalline free energy difference; related to kinetic versus thermodynamic control of crystal growth.

Formulas

Barandiaran-Colmenero critical cooling rate, Hoffman equation, temperature of maximum crystal growth rate.

How it is measured or obtained

As Tm minus the crystallization peak temperature on cooling at a given rate baird2010 P2; baird2013 R1; as the temperature scale for growth data trasi2014a P5; in dispersions it shrinks when the polymer depresses the drug's melting point zhu2011 C20. In the PEG dispersions it was not measured: the supercooling is inferred from the liquidus of the drug–PEG phase diagram zhu2012 C32; zhu2011 R1. The 25 versus 40 °C solidification of pure PEG was used as a control of how much a change of PEG supercooling moves the lamellar long period (about 1 nm) zhu2011 R8, C21. In the amorphous solubility papers Tm comes from DSC and T∞ from the enthalpy of conversion with ΔH(c → sl) set to zero in the heat-capacity cycle almeidaesousa2015 E9, R16. In the polymer growth papers, from the onset melting temperature of 10% physical mixtures at 10 °C/min trasi2012a P5, R9.

Values in the library
materialvalue or classconditionspaper ids
caffeine3.0 °CDSC cooling at 20 °C/minbaird2010 R12.j
tolbutamide80.8 °CDSC cooling at 20 °C/minbaird2010 R12.v
4-biphenylmethanol; haloperidol15.1; 90.8 °Crapid cooling at 500 °C/minbaird2013 R1.a, R1.f
class I-A meltscrystallized after about 5–35 °Crheometerbaird2012 R4
vemurafenibabout 25 °C before crystallizationDSC cooling at 10 °C/minlu2016 R1
benzocaine in PEG 20:80melting point 55 against 92 °C for the pure drugdispersionzhu2011 C20
felodipine growth runsΔT 71.4–31.4 °C over 70–110 °Cmelt growthkestur2010b R17
benzocaine in PEG 3350supercooling at 25 °C: 89 − 25 = 64 K neat against 48 − 25 = 23 K at the eutectic (synthesis)cited melting pointszhu2012 M3, C32
PEG 3350 with 20 wt% benzocaine; ibuprofen; fenofibrateliquidus about 54.5, 57, 58.0 °C against about 60.5 °C for PEG, a depression of 6.0, 3.5, 2.5 K (synthesis)read from the phase diagramszhu2011 R1.g, R2.g, R1.a; zhu2012 R24.d
pure PEG 335025 °C: t_{50} about 23 s; 40 °C: about 48–64 ssynchrotron XRDzhu2012 R4, R16
ibuprofen, phenylbutazone, tolnaftateTm − T 51, 82, 86 K; Tg − T∞ 81, 42, 143 K25 °Calmeidaesousa2015 R16
diclofenac, sorafenib, griseofulvinTm − T 147, 180, 192 K; Tg − T∞ 145, 107, 122 K25 °Calmeidaesousa2015 R16
telaprevirTm 238 °C against 37 °C, a gap of about 200 KHoffman estimate 2.7 times the onsetmosqueragiraldo2015 M1, C23, C24
CRH1Tm 141.7 °C, so Tm − T = 116.7 K (synthesis: 141.7 − 25)all three free-energy routes agree within 1.3 µg/mL[almeidaesousa2016 M1, R2]
acetaminophen with 10% w/w of seven polymersTm 166.7–168.6 against 169 °C; supercooling similar for allDSCtrasi2012a R9, C19
flutamide with 10 mol% polymersTm 108–110 °C; supercooling hardly changedDSCtrasi2012b R7, C33
celecoxib with TPGS and PVPTm 151.49 °C, 12 °C below the drugDSCmosqueragiraldo2014 R5.f, C18
six compounds with 1–20% PHBTm of physical mixtures 0–4 °C lower, so supercooling taken as unchangedDSCsato2016 R7, C18
indomethacin δ, α, γTm 127 (cited), 152 and 160 °CDSCtian2017 M1
Pitfalls and disagreements

Undercooling did not always rise with cooling rate and scattered between runs baird2010 R12, C18. Hoffman estimates lose accuracy at large undercooling almeidaesousa2015 C34; mosqueragiraldo2015 C24. Low polymer levels barely shift Tm, so the supercooling is similar with and without polymer trasi2012a C19; sato2016 C18. In the PEG dispersions the supercooling and the mobility of benzocaine were not measured, and the two cited melting points differ between the two PEG papers (89 and 48 °C against 92 and 55 °C for the 20/80 blend) zhu2012 C32, C33; zhu2011 R1. The authors do not weigh reduced PEG supercooling as a cause of slower PEG crystallization, only hydrogen bonding zhu2012 C44. In the amorphous solubility papers Table 10 does not predict the size of the Hoffman error drug by drug: sorafenib has 180 K undercooling and equal Hoffman and Cp ratios almeidaesousa2015 X8. Small melting-point shifts were used to rule the driving force out as the cause of growth differences between polymers trasi2012a C19; trasi2012b C33.

← Concepts and formulas

concept · as used in Crystallization tendency

desupersaturation

The fall of the dissolved concentration after supersaturation was reached, because crystals nucleate or seeds grow; in an ASD it "corresponds to the crystal growth rate exceeding the dissolution rate" [moseson2020a C14]. A plateau is not the same as a fall: a crystal-free bicalutamide dispersion that stayed at about 18 µg/mL (30 µg/mL dose) without desupersaturation was read as crystallisation inside the dissolving solid, which caps the concentration without lowering it afterwards [moseson2021 C17, R14]. In the posaconazole pH-shift test the concentration falls below the LLPS concentration (about 19 µg/mL in the intestinal medium) when crystals present or newly formed consume the supersaturation; this is not a return to the crystalline solubility of 1.2 µg/mL [shah2024 R11, X15]. New aspect (Bioperformance): the loss of supersaturation, mostly by crystallization from solution or in the matrix; its onset is read from a concentration fall [elkhabaz2017 C19, C23]. In dissolution of enzalutamide–PVPVA dispersions and tacrolimus capsules the fall after the maximum was read as crystallization, from seeds in the product or formed in the medium; absorption lowers the donor too [wilson2018 C18; purohit2019 C17, C34; hate2019 C20, C30]. New aspect (Crystallization from solution): with seed crystals present it reflects growth of the seeds; without seeds the crystals must first nucleate, so the profile has a flat start (the induction time) followed by a fall [alonzo2012 P4, P5; ilevbare2013c R7]. New aspect (Supersaturation from ASD dissolution): the fall of the concentration after a dissolving amorphous solid or dispersion has supersaturated the medium, read as crystallization from solution, sometimes together with crystallization of the remaining solid [alonzo2010 C16, C21; jackson2015 C14].

Findings

ASD process: F25, F26, F27, F28, F38, F39, F40, F41, F46. Bioperformance: F9, F10, F11, F12, F14, F16, F23, F28, F29, F30. Crystallization from solution: F4, F5, F14, F20, F23, F24, F27, F33, F42. Supersaturation from ASD dissolution: F4, F5, F6, F15, F16, F20, F22, F26, F28.

Related concepts

read in non-sink dissolution; driven by crystal seeding; slowed by crystal growth inhibition; contrasts with matrix crystallization. Loss of supersaturation; caused by solution crystallization and secondary nucleation; also lowered by an absorptive sink; driven by crystal growth rate and nucleation induction time; summarized by supersaturation factor; related to residual crystallinity.

Formulas

power-law crystal growth rate, initial growth rate from the desupersaturation slope, supersaturation factor, supersaturation ratio, overall mass crystal growth rate.

How it is measured or obtained

From the concentration–time profile after the maximum, or from seeded runs: 50 µg/mL indomethacin made by antisolvent addition and 10–50 µg/mL bulk seeds added, with the growth rate R_G in µg/s taken from the profile (extraction method not printed) [moseson2020a P10, P20, X12]. In the Bioperformance papers: concentration–time profiles, UV at 233 and 350 nm for induction time, with SHG or PLM to confirm crystals [elkhabaz2017 P5, P8, P10]. In the crystallization-from-solution papers: concentration against time by a fibre-optic UV probe on second-derivative spectra (every 2–5 s), in seeded runs (slope of the first 2–5 min as initial growth rate) and unseeded runs (flat start, then fall); light-scattering extinction and pyrene fluorescence give the matching signals for droplets and crystals [alonzo2012 P3; ilevbare2012a P8; jackson2014 R15]. In the ASD dissolution papers: in situ UV during non-sink dissolution or after a solvent shift; a rise of the pyrene ratio confirms loss of the drug-rich phase [konno2008a P5; alonzo2010 P3, P4; jackson2015 P2, P3]; drug in the filtered liquid of suspensions [chen2015a P11].

Values in the library
materialvalue or classconditionspaper ids
IDM, 50 µg/mL, no polymerunseeded about 50 → 25.6 µg/mL at 120 min; with 50 µg/mL seeds about 26.9pH 4.5, 37 °C[moseson2020a R3.a, R3.d]
IDM, 50 µg/mL, 5 µg/mL PVPVA, 50 µg/mL seedsabout 50 → 37.1 µg/mL in 720 minsame[moseson2020a R3.h, X6]
IDM–PVPVA extrudatesslight fall after the peak above 3% crystallinity (131-2 about 40.6 → 39.4; 117-2 about 30.9 → 28.8)50 µg/mL total dose[moseson2020a R6]
BCL–PVPVA S/155 and M/155about 32.4 → 24.5 and about 28.2 → 16.2 µg/mL150 µg/mL dose, 4 h[moseson2021 R12]
paclitaxel–HPMCAS with 18 wt% seeds (outside)desupersaturation by crystal growth; HPMCAS cannot block it completely16 h[que2018 C30]
tacrolimus capsules (outside)steep decline after the maximum shows that seeds grow rapidly in 100 mLpH 4.5[purohit2018 C43]
PCZ–HPMCAS 10% DL, 0.3% crystals against nonepeak about 178 → 18.5 µg/mL at 150 min against about 172.5 → 125 µg/mL at 300 minpH shift at 60 min, dose about 200 µg/mL[shah2024 R12, R14]
PCZ–HPMCAS 25% DL, 3.2% crystals against nonecrystallization after 45 min against about 1.5 hsame[shah2024 R15, C29]
BCL–PVPVA 30% DL, 3% SDS, cryomilled 5 minpeak about 84% at 45–60 min, then about 78% (about 23 µg/mL)dose of at most 30 µg/mLmoseson2022b R10
PCZ 20 µg/mL in FaSSIF V2, HPMCAS 0 / 9 times the drug; seeds 0 / 5 / 10 µg/mL% left at 30 min about 66 / 46.5 / 25; about 99.5 / 96 / 74.5seeded solution[shah2024 R22]
ezetimibe PVP ASD, FeSSIF195 µg/mL at 15 min to 41 at 30 min37 °C[elkhabaz2017 R14]
ezetimibe ASDs, bufferdecline after about 30 min[elkhabaz2017 R10, C19]
tacrolimus generic, 100 mLabout 98% released at 20–30 min, 39.5% at 120 minmedium I[purohit2018 R7]
felodipine, no polymer, unseededabout 5.35 → 1.54 µg/mL from about 10–15 min to 150 minS 10[alonzo2012 R15]
paclitaxel–HPMCASnone unseeded; present with 18 wt% seeds16 h[que2018 R13]
pazopanib19.8 to about 1 µg/mL at 60 minpH 6.5[sugihara2018 R8]
enzalutamide PVPVA ASDsmaximum close to the amorphous solubility, then fall; 10:90 HPMCAS peak only 10 µg/mL with crystals in the medium100 µg/mL total, 12.5 mL, pH 6.5[wilson2018 R6, C18]
tacrolimus, seeded solutionsrate about 0.39 at 50 µg/mL against 0.075 at 20 µg/mL; growth order 1.677seeds from the crystallized generic[hate2019 R17, R18, C30]
tacrolimus 50% and 100% crystalline capsule contents15 and 4 µg/mL at 2 h, still above the crystalline solubility5 mg in 100 mL, pH 4.5[purohit2019 R4]
felodipine, 1 µg/mL HPMC, seeded; unseeded5.1 → 2.17 µg/mL in 58 min; 5.38 → 5.03 over 198 minS 10, 25 °C[alonzo2012 R18]
ritonavir seeded, none; CAP Adp 0.8510.0 → 6.9; 10.15 → 9.25 µg/mL in 2 minS 7.6, 37 °C[ilevbare2012c R7]
danazol 20 µg/mL, no polymerapparent UV concentration 19.2 → 14.4 → 11.8 → 2.1 → 1.4 µg/mL at 1, 5, 10, 30, 60 min25 °C[jackson2014 R15]
celecoxib, HPMC, start S 14.7S 15 for 15 min, 7.8 at 35 min, 4.9 at 100 min, 4.3 at 200–240 min37 °Cilevbare2013c R16
paclitaxel with 18 wt% seeds (ASD dissolution chapter)desupersaturation immediate without polymer37 °C[que2018 R7]
telaprevir seeds, 1.86 mM STCDCflat at about 11.2 and 72 µg/mL for 240 min; buffer 4 and 6δ 2.4 and 12.5[lu2017a R11]
telaprevir after 15 h, δ 12.5residual 60.5 (STCDC), 91.7 (SGCDC), 9.0 (STC) of 65.2–95.2 µg/mLdiffusion cell[lu2017a R12]
acetaminophen seeded19.5–19.8 → 13.0 (none) and 16.1 mg/mL (HPMC) at 24 hS 1.5, 20 µg/mL polymertrasi2015 R11
atazanavir, posaconazole after nucleation, three mediaatazanavir 1.0, 4.2, 0.3; posaconazole 0.6, 1.9, 1.7 µg/mL·min (buffer-V1, FaSSIF-V1, composite)37 °C, slope after the break[elkhabaz2021 R8, E7]
posaconazole at S 33.6 with nanocrystal seedsfrom about 8.5 to 4.9 µg/mL by 4 min (2 µg/mL seeds), to about 3.4 within about 1 min (20 µg/mL)PBS pH 6.5duong2022b R13
felodipine 50 µg/mL nominal, 150 µg/mL PVP; none≈6.3 → ≈1.9 µg/mL in 240 min; ≈2.7 → ≈1.05solvent shift, 37 °C[konno2008a R12]
indomethacin, solvent shift, no polymer≈27.3 µg/mL held about 25 min, then ≈5.3 at 90 minpH 2, 25 °Calonzo2010 R18
felodipine 90:10 HPMC, 90 µg/mL addedapparent ≈75 µg/mL at 5 min, ≈22.6 at 50 min37 °C[alonzo2011 R2]
danazol 90:10 PVP dispersion≈9.2 µg/mL at 60 min to ≈2.2 at 420 min, pyrene ratio risingpH 6.8, 25 °C[jackson2015 R12]
nifedipine–HPMC 50:50≈100 µg/mL at ≈22 min to ≈41 at 150 min37 °Craina2014b R14
celecoxib–HPMCAS suspension, SDSS ≈8.6 at about 5 min to ≈3.2 at 1 hpH 3, 25 °C[chen2015a R17]
celecoxib 22 µg/mL: none / 5 µg/mL PVP-K12 / PVP-K29/32≈20.8 → ≈4.1 in 1 h / ≈20.9 → ≈4.4 by 8 h / ≈9.1 µg/mL at 8 hsolvent shift, pH 6.8, 37 °Cxie2015 R2
celecoxib–PVP-K12 50% dispersion≈21.0 µg/mL near 6 h → ≈6.5 at 16 hpH 6.8, 37 °Cxie2015 R6
celecoxib 22 µg/mL seeded: none / 5 µg/mL HPMCAS≈22.5 → ≈4.0 in 30 min / → ≈16.0 µg/mL at 4 hpH 6.8, 37 °C[xie2016a R11]
tacrolimus 50 µg/mL: none / 50 µg/mL HPMCfall from ≈150 min / from ≈400 minpH 4.5, 37 °Ctrasi2016 R9
indomethacin–EPO 3:7 dispersion≈77 µg/mL (UV) → 8.8 µg/mL (HPLC) at 1 hpH 2.2, 37 °C[xie2017 R11]
lopinavir 50%: EUD, CAP / HPMCPloss within ≈10–12 h / none in 48 hpH 3, 25 °C[li2018 R6]
paclitaxel–HPMCAS 49/51 with 18 wt% seeds≈31–33 µg/mL reached, then ≈7.9–25.3 µg/mL left at 16 h depending on the seed typepH 6.8, 37 °C[que2018 R14]
posaconazole–HPMCAS 10% dispersion, high against low dose≈22–24 µg/mL to 120 min then ≈6–8; ≈17–21 µg/mL to 90 min then ≈8 µg/mLFaSSGF → FaSSIF, 37 °C[elkhabaz2019a R6, R12]
praziquantel–HPMC E5 LV dispersions, 1.5 mg/mLa 30% drop in FaSSIF after about 75–82%; R-enantiomer lost dissolved drug in every medium37 °C[alvarenga2025 R14, C36]
Pitfalls and disagreements

Growth rates are vessel mass rates, not normalised to seed area [moseson2020a X12]. In the bicalutamide tests the desupersaturation at 150 µg/mL is attributed to growth from solution, whereas the overall low plateau is attributed to growth in the solid [moseson2021 C21, C46, X4]. The IDM extrudates may have lost concentration by growth into the surrounding amorphous solid rather than from solution [moseson2020a X15]. In the posaconazole test the focused beam reflectance signature read as secondary nucleation for the crystal-containing 25% dispersion has the same form as the one read as primary nucleation for the crystal-free one [shah2024 K3, K4, X14]. For bicalutamide with cryomilled surfactant powders the loss is attributed to nuclei in the milled matrix and was not measured moseson2022b C26. In the Bioperformance papers concentration alone cannot separate LLPS from crystallization in turbid media [elkhabaz2017 C30]. Absorption also lowers the donor (F14). With an absorptive sink the fresh generic tacrolimus did not lose supersaturation at 240, 100 or 40 mL, unlike in a closed vessel; only at 40 mL did total dissolved drug fall for the 20% sample (F14, F28) [hate2019 R11, C20, C35]. Apparent UV concentration of a colloidal solution is distorted by droplet size changes: at 20 µg/mL danazol the early fall is attributed to droplet coalescence, not crystallization, and orthogonal methods are needed [jackson2014 C29, C54, C55]. The seeded profile includes any secondary nucleation [ilevbare2012b X18]. After crystallization with HPMC the celecoxib solution sat at S 4.3, not 1, at 240 min ilevbare2013c R16. In the ASD dissolution papers apparent UV falls above the amorphous solubility can come from particle growth or settling as well as from crystallization [alonzo2011 C14; jackson2015 C12]. With HPMC the felodipine solution kept falling slowly; the PVP and buffer solutions approached but did not reach the crystalline solubility in 4 h [konno2008a X10]. After crystallization the celecoxib readings settled at ≈2.6–2.9 µg/mL, above the 1.5 µg/mL crystalline solubility measured by HPLC [xie2016a X4]. With crystals present from the start, the onset of desupersaturation marks the time at which growth outpaces redissolution of amorphous material, not the onset of crystallization [elkhabaz2019a C49].

← Concepts and formulas

concept · as used in Crystallization tendency

diffusion-controlled crystal growth

Crystal growth whose rate is limited by transport of molecules to the crystal surface rather than by their incorporation into the lattice. From the supercooled melt it shows as growth that slows on cooling although the driving force rises, because transport (viscosity) dominates kestur2010b C22, C23. In solution growth an overall growth order of 1 is read as diffusion control and 2 as integration control [alonzo2012 C3].

Findings

F3, F19, F28.

Related concepts

part of crystal growth rate; depends on molecular mobility; contrasts with integration-controlled growth; contrasts with glass-crystal growth mode.

Formulas

the two-dimensional nucleation growth rate printed by the melt paper is listed on the chapter page as table only kestur2010b E5; Stokes-Einstein equation gives the polymer diffusivity used with it.

How it is measured or obtained

From the temperature dependence of melt growth rates compared with the growth equation v = (k/\eta)\exp\left(-\Delta G^{*}/k_B T\right) kestur2010b E2, R7; in solution from the slope of ln growth rate against ln supersaturation [alonzo2012 C3].

Values in the library
materialvalue or classconditionspaper ids
felodipine form II with and without PVPgrowth falls on cooling over 70–110 °C (ΔT 71.4–31.4 °C): transport-limitedmelt filmskestur2010b R13, R17, C23
felodipine in aqueous solution (outside)HPMC shifts growth toward integration controlseeded desupersaturation[alonzo2012 C33]
diffusion-controlled growth against the glass-crystal mode0.1–1 against 1700–2400 molecular layers per relaxation time (cited); felodipine form II below 1felodipine at 45 °Ckestur2013a R16, R18
six compounds with and without PHBall growth temperatures below 0.94 Tm, so growth taken as transport-limited ("is expected to be")melt filmssato2016 E2, C15
low-Tg compounds with PHBretardation attributed to PHB chains impeding transport to the crystal ("most likely")melt filmssato2016 C25

Evidence table: diffusion-controlled crystal growth.

Pitfalls and disagreements

The supercooling of the felodipine melt study is computed from the form I melting point although form II grew, so the stated ΔT range is too large kestur2010b X15. Below the temperature of maximum growth, transport limitation is the expected regime for any melt; it does not by itself identify the polymer mechanism kestur2010b C23.

← Concepts and formulas

concept · as used in Crystallization tendency

drug-polymer interactions

The contacts between drug and polymer in an amorphous dispersion, relative to the contacts in the pure components: strong adhesive (drug–polymer) interactions formed at the expense of weak cohesive (self) interactions make mixing exothermic, adhesive interactions that require breaking strong cohesive ones do not marsac2009 C3, C36. The interactions are hydrogen bonds where the drug has a donor and the polymer an acceptor, and, for a drug without donors, perhaps dipole–dipole forces and van der Waals contacts [taylor1997 C35; marsac2009 C41; rumondor2009a C23]. Specific interactions have been postulated as one way polymers stabilize amorphous drugs, and as a condition of single-phase dispersions kestur2010a C3; marsac2009 C3. In this chapter they are the common thread of the miscibility papers (whether the pair mixes) and the inhibition papers (whether the polymer slows crystallization). The later papers add ionic (acid–base) contacts of E100, enteric polymers and PSSA with basic or acidic drugs [wegiel2013a C16, C35; hiew2022a C25, C48; vaneerdenbrugh2011a C22] and a weak halogen bond between compounds with Cl, Br or I but no donor and the PVP carbonyl [que2020 C1, C25]. A polymer can also bond to itself (PAA, dextran), which competes with the drug [rumondor2009a C45; vaneerdenbrugh2012a C36].

Findings

F1, F2, F3, F4, F7, F8, F12, F13, F14, F15, F17, F20, F21, F23, F24, F25, F26.

Related concepts

includes hydrogen bonding and dipole-dipole interactions; measured by Flory-Huggins interaction parameter and activity coefficient; determines drug-polymer miscibility and crystal growth inhibition by polymers; related to enthalpy of mixing; includes acid-base interaction and halogen bonding.

Formulas

Flory-Huggins melting point depression, Flory-Huggins activity equation.

How it is measured or obtained

Spectroscopically, by shifts of IR bands of the drug and the polymer carbonyl in dispersions against physical mixes and amorphous references taylor1997 C24, C26; marsac2009 P4; by a second principal component of the IR spectra beyond calculated physical mixtures [rumondor2009a P5, C9]; thermodynamically, by the melting point depression of the crystalline drug in the polymer (χ) and by the solubility of the drug in a liquid analogue of the polymer (activity coefficient) marsac2009 P6, P7; by DSC and PXRD for the resulting phase state [rumondor2009a P2, P8, P9]. Interactions are not measured at the temperature and polymer level of the growth studies kestur2010a X5. Further methods: IR band shifts of the polyphenol OH and the polymer C=O in films wegiel2013a P4; XPS of N 1s and of halogen core levels [hiew2022a P16; que2020 P5]; solution NMR titration with a monomer analogue for a binding constant [que2020 P6]; crystal-structure statistics of competing groups [vaneerdenbrugh2011a P5].

Values in the library
materialvalue or classconditionspaper ids
indomethacin–PVPacid OH bonded to PVP carbonyl; dimers opened; incomplete bindingIR, 5–70 wt% PVP; physical mixes show nonetaylor1997 C35, C26, C37
felodipine–PVP, nifedipine–PVPone phase over 10–90% and 25–70% PVP; hydrogen bonding "presumably" behind the second PCIR, PXRD[rumondor2009a C38, C20]
felodipine–PAAlittle mixing at 30–70% PAA; competition from PAA self-association "speculated"DSC, IR, PXRD[rumondor2009a C44, C45]
ketoconazole–PVPnear-athermal mixing, no carbonyl shift; dipole–dipole forces "speculated"melting, FTIRmarsac2009 C40, C41, R12.h
felodipine with PVP, PVP/VA, HPMCAS, PVAcstrength PVP > PVP/VA > HPMCAS ≥ PVAcNH shifts at 50 wt% polymerkestur2010a C26, R9
nimesulide with PVP, PVP/VA, PVAcpresent but small; limited by the intramolecular bondIR, 20–50 wt% polymerkestur2011 C23, C24, C27
amide and basic drugs with PAA, PSSAhighest crystallization resistance; acidic drugs lowestspin-coated films, 7 daysvaneerdenbrugh2010b C23, C24, C27
indomethacin–PVP K12χ −0.82 (melting), −1.83 (EP solubility)most favourable of five pairsmarsac2009 R7.e, R9.e, C42
naringenin, quercetin with PVP; with E100hydrogen bonding strongest with PVP; acid–base contact with E100 (amine bands lost at 50% quercetin)IR, filmswegiel2013a R2, R4, R6
lumefantrine with enteric polymers and PVPVAprotonated 19–55% with HPMCAS, HPMCP, CAP, Eudragit L 100; 0% with PVPVAXPS, 25% drug loadinghiew2022a R20
clotrimazole, loratadine, brotrimazole, Me-DIBF with PVPVAhalogen bond: K_F 0.56–1.11 M⁻¹, carbonyl shift 2–6 cm⁻¹NMR, IR[que2020 R12, R11]
15 drugs, 7 polymerscomplementary donor and acceptor groups give the highest amorphicity indexspin-coated films[vaneerdenbrugh2011a R3, C20, C33]

Evidence table: drug-polymer interactions.

Pitfalls and disagreements

In every inhibition study cited here the interaction is inferred or measured under other conditions than the crystallization test, and its causal role is argued, not demonstrated kestur2010a X5, X6; kestur2011 X9, X10; vaneerdenbrugh2010b C33. The two thermodynamic routes agree in sign but not in size marsac2009 X7. A strong donor polymer is not enough when it bonds to itself (PAA with felodipine) or when the drug donor is tied up intramolecularly (nimesulide) [rumondor2009a C45; kestur2011 C27]. Interaction does not always decide the outcome: PAA, a strong donor, left flutamide growth almost unchanged trasi2012b R3, C7. The strength of the ionic and halogen contacts was not measured in the dispersions, and their evidence rests on band loss, binding-energy shifts or spectra that are not shown [wegiel2013a X10; que2020 X11, X12; vaneerdenbrugh2011a X13]. The ionic contact that protects against crystallization also slows the dissolution of the enteric polymers hiew2022a C53, C65. A complementary polymer did not always protect: the IR ranking of the polyphenol polymers held at the extremes only wegiel2013a X4.

← Concepts and formulas

concept · as used in Crystallization tendency

drug-rich nanodroplets

The dispersed drug-rich phase formed by LLPS, usually of 100–300 nm, which dissolves fast and is expected to replenish absorbed drug [indulkar2016 C4, C25]. Droplets are too large to cross the 3–4 nm (regenerated cellulose, 12–14 kDa) or 15 nm (hollow fibre) pores of the membranes used [indulkar2016 C11; hate2018 C52]. In enzalutamide dispersions the same species is called amorphous aggregates, about 100 nm; in simulated and pooled human intestinal fluids spheres of 0.1–0.5 µm were found [wilson2018 R11, C45; elkhabaz2019b R4, C19, C24]. New aspect (Supersaturation from ASD dissolution): fast-releasing dispersions put drug above the amorphous solubility into a submicron colloid within minutes, and additives decide its size and how soon it crystallizes [ilevbare2013d C59, C60; jackson2015 C29]. New aspect (Release mechanism and LoC): from dissolving dispersions, nanodroplets formed only where release was congruent or close to congruent, so that the drug concentration exceeded the amorphous solubility [purohit2017b C52; indulkar2019a C5; saboo2019 C30]. The 2017 isotope study concludes that they form from drug that has first dissolved, followed by LLPS ("strongly suggest"), not by release of drug-rich domains from the hydrated matrix [indulkar2017b C26]; Purohit and Taylor reach the same reading for ritonavir films ("likely") [purohit2017b C57]. When drug release is controlled by polymer dissolution, colloid formation is "highly dependent" on drug loading [que2019 C55]. Their size and life after release depend on the drug, the polymer and small amounts of surfactant: HPMCAS kept elbasvir and loratadine droplets small, and SDS, TPGS or Tween 80 at 4–50 µg/mL kept clopidogrel droplets stable for 4 h while CTAB agglomerated them [yang2021 C4; correasoto2022a C12, C38]. Built into ritonavir–PVPVA at 30%, the Spans kept the released droplets near 1 µm, while TPGS and Tween 80 let them grow to several micrometres, which the authors suggest "could possibly" come from these surfactants partitioning into the droplets and lowering their Tg indulkar2022 C47, C48. In dissolving ritonavir–PVPVA compacts at 10 and 20% droplets formed near the gel–solution interface and entered the solution with the dissolving polymer, which the authors explain by a water content needed to cross the binodal [yang2022a C34, C35, R10, R11].

Findings

Bioperformance: F5, F8, F15, F19, F22, F23. Supersaturation from ASD dissolution: F8, F9, F12, F13. Release mechanism and LoC: F4, F5, F12, F13, F16, F25, F27.

Related concepts

produced by liquid-liquid phase separation; act as nanodroplet reservoir effect; part of the drug-rich phase; related to droplet coalescence and particle absorption of light; form under congruent release above the amorphous solubility. Coarsen by drug-rich phase coarsening.

Formulas

Stokes-Einstein equation.

How it is measured or obtained

NTA (scattering or fluorescence mode with Nile red), DLS, the 369 nm band of atazanavir [indulkar2016 P4; hate2018 P3, P4, P6]. Nile Red fluorescence of the pellet against aggregate mass, and DLS [wilson2018 R8, R11]; SEM of a centrifuged pellet with the drug's own fluorescence in human fluid [elkhabaz2019b R3, R4]; Z-average by DLS under agitation [ueda2020a R5, R6]; size and zeta potential with polymer or surfactant additives [hate2022 R17]. In the ASD dissolution papers: light scattering in a flow-through cell, nanoparticle tracking of samples or 0.45 µm filtrates, SEM of dried samples [ilevbare2013d P3, P8; jackson2015 P5; trasi2015b P8]. In the release-mechanism papers: NTA, DLS, visible turbidity, environment-sensitive probes in solution and free drug after ultracentrifugation [indulkar2017b R6, R7; purohit2017b R7, R8; indulkar2019a R9; que2019 R5, R12; saboo2019 R11, R12].

Values in the library
materialvalue or classconditionspaper ids
clotrimazole111 / 223 / 246 / 257 nm at 8 / 10 / 15 / 20 µg/mL; 1.2 × 10⁸ to 12.5 × 10⁸ per mLpH 8, NTA[indulkar2016 R4]
atazanavir, solvent shift204–226 nm, PDI 0.08–0.15100–500 µg/mL, DLS[hate2018 R6]
atazanavir 90:10 HPMCAS and HPMC ASDs294–326 nmclosed and absorptive[hate2018 R12]
atazanavir with PVPVAabout 245 nm rising to about 700 nm at 75 min and 10 µm at 105 minagglomeration[hate2018 R20]
telaprevir 90% HPMCAS ASDabout 300 nmDLS (data not shown)[mosqueragiraldo2018b R3]
ritonavir ASD (Tres 2018)696 / 294 nm (ritonavir / lopinavir)NTA[tres2018 R18]
enzalutamide 10:90 and 50:50 copovidone ASDsabout 100 nm; about 50 against about 3 µg/mL of 100 as aggregatespH 6.5, 1 h dissolution[wilson2018 R10, R11]
atazanavir, posaconazole in pooled human fluidspheres of 0.1–0.5 µm, no needlesSEM of pellet, 100 µg/mL HPMC-AS[elkhabaz2019b R4]
atazanavir, clozapine with HPMCAS-LF and -HFAS-LF droplets below 300 nm over 3 h; AS-HF kept the initial size; clozapine about 100 nm (AS-HF) against 250 (AS-LF)300 rpm, 37 °C[ueda2020a R6]
atazanavir 150 µg/mL with polymer or surfactant additivesdroplets of about 280 and about 1000 nm gave the same mass flowPAMPA, 2% SDS receiver[hate2022 R17]
ritonavir–PVP dispersion at 20 µg/mLabout 350 nm at onset; ≈5590 nm by 120 min100 mM buffer, 37 °C[ilevbare2013d R1, R5]
ritonavir 20 µg/mL with 5 µg/mL CAAdP 0.33210−214 nm before crystallizationsolvent shift[ilevbare2013d R12]
danazol 90:10 dispersions246–284 nm25 °C[jackson2015 R6]
ritonavir; ritonavir + lopinavir, HPMCAS dispersionsabout 500 nm; about 160 nm in 0.45 µm filtrates1 h[trasi2015b R7]
felodipine 90:10 HPMC dispersion≈185 → ≈300 nm, read as crystalline25 °C[alonzo2011 R8, C33]
nifedipine 10, 20% with HPMC-E5 or PVPVA≈190–222 nmNTA, 5 °C, D2O[indulkar2017b R7]
ritonavir 10:90 films with PVPVA, HPMCAS, PVP200, 250, 315 nmNTA, fresh films[purohit2017b R8]
ritonavir–PVPVA 10% tablet218 ± 15 nm; media of 10–25% turbidDLS at 30 min[indulkar2019a R9]
ledipasvir–PVPVA 2, 5%cloudy medium; ≈225–257 nm at 5%DLS[que2019 R5, R12]
nilvadipine 5, 10%; cilnidipine 5, 10, 15%237 ± 8, 246 ± 3; 255 ± 1, 300 ± 2, 366 ± 7 nm; none at higher loadingsDLS, about 1 h[saboo2019 R11]
indomethacin–PVPVA 5, 10%250–350 nmNTA, 30 minsaboo2020 R7
felodipine with 5% TPGSZ-average ≈237–279 nmDLS, 60 min[saboo2021 R17]
elbasvir 2.5%; loratadine 10%PVPVA ≈670 → 1190; ≈820 → 2120 nm; HPMCAS ≈280 → 390; ≈420 → 570 nmDLS, 60 min[yang2021 R3, R6]
clopidogrel≈520 nm in buffer; stable with SDS, TPGS or Tween 80; ≈12-fold growth with CTABDLS, 4 h[correasoto2022a R5, R6]
four phenolphthalein analogues179–364 nmDLS, 30 min[que2021 R22]
ritonavir–PVPVA 30:70:5 with Span 85; Span 20; none; SDS; TPGS; Tween 80≈330–960; ≈550–1410; ≈940–2200; ≈2400; ≈3840 nm; ≈10 µmDLS of the medium during releaseindulkar2022 R11, R13
plasticized dispersions (systems not specified)500–1500 nm after 90 minDLS[correasoto2023 R5]
MePHPH–PVPVA 5–30%; PHPH–PVPVA 5%nanoparticles formed within the gel; onset 601 ± 68 µm beyond the 356 ± 18 µm gelin situ confocal[deac2023a C27, R8, C31]
drug-rich phase released from 30% MePHPH; from 5% PHPH98.8 against 26.2 wt% drug; Tg 16.8 against −5.8 °CHPLC, DSC[deac2023a R12]
PHPH; MePHPH nanoparticles with PVPVAPVPVA 71.4–78.0; 0.5–7.5 wt%37 °C, two preparation routes[deac2023b R2]
Pitfalls and disagreements

The size is quoted as 110, 200, 220 or 250 nm in one paper [indulkar2016 X9]. Size depends on polymer and agglomeration [hate2018 C26, C27]. Counts stop at 150 µg/mL in the atazanavir study [hate2018 X10]. The enzalutamide aggregate amounts come from a Nile Red calibration, and the aggregate-rich ASD also dissolved fastest and left less solid in the dosed suspension (F23) [wilson2018 R8, X13, X14]. Size fell with polymer concentration and was smaller with AS-HF (F19) [ueda2020a C18]; in a PAMPA cell the polymer or surfactant type, not the size, shaped the transport gain (F22) [hate2022 R17]. Droplets pass 0.45 µm filters [trasi2015b C22]. Sizes depend on method (SEM of dried samples against DLS) and medium [ilevbare2013d X12, R9]. Whether the felodipine particles were droplets or crystals was not measured directly [alonzo2011 X7]. Nanodroplets of ritonavir tablets at 20–30% are inferred from turbidity and concentration, not sized [indulkar2019a X5]. The link between congruency and droplets partly follows from the amount of drug released by the sampling time [saboo2019 X14]. The larger cilnidipine droplets at higher loading "could indicate" lower stability or a different formation route [saboo2019 C19]. The slow-exchange premise of the isotope test was shown with solvent-made droplets only [indulkar2017b X5].

← Concepts and formulas

concept · as used in Crystallization tendency

drug-rich surface layer

A polymer-depleted, drug-enriched layer at the surface of a dissolving ASD, left when the polymer dissolves ahead of the drug above the limit of congruency. Moseson (2022) shows it on bicalutamide–PVPVA compacts and says that this layer, hydrated and mobile, crystallizes within 20–30 min, which "could prevent release, promote surface crystallization, and prevent or slow continued hydration" moseson2022b C46, C43. In the 2021 paper it was inferred from the shape of the PVPVA release curve and pitted surfaces, "presumably drug-rich" [moseson2021 C32, C58]. It is the general case of surface drug enrichment applied to dissolution. New aspect (Release mechanism and LoC): the layer is the drug-controlled surface of the older two-component literature (sulfathiazole–PVP, cited) [purohit2017b R23], and the nifedipine study states that if water-induced phase separation forms such a layer, the drug controls release and the carrier effect of the polymer is lost [indulkar2017b C33]. For ritonavir the authors add that drug molecules "can" move laterally on the dissolving surface and that drug-rich nanodomains "can" coalesce into contiguous regions, ritonavir being a supercooled liquid at 37 °C [indulkar2019a C28, C40]. Saboo and co-workers (2021) model it as what a contiguous drug-rich network leaves once the polymer-rich phase dissolves ("likely") [saboo2021 C39]; for clopidogrel, faster polymer loss leaves a drug-enriched compact that "likely accounts for" incomplete release [correasoto2022a C25].

Findings

ASD process: F27, F38, F48. Release mechanism and LoC: F3, F7, F20.

Related concepts

forms above the limit of congruency during incongruent release; sits on the gel layer; leads to matrix crystallization and surface crystallization. Its barrier role is the drug-rich barrier layer; sets drug-controlled release; mobile where the drug is a supercooled liquid.

Formulas

None.

How it is measured or obtained

SEM-EDX of exposed compacts: fluorine occurs only in the drug, so F/F0 above 1 means drug enrichment; confocal fluorescence with a drug-rich dye; SEM of the surface for crystals moseson2022b P14, P15, P16, P17, C8. In the release-mechanism papers: IR of the tablet surface, micro-CT and SEM of dried compacts [indulkar2019a P7, P8; que2019 P5; saboo2019 P14].

Values in the library
materialvalue or classconditionspaper ids
30% BCL–PVPVA compact surfaceF/F0 = 1.23 ± 0.07 (about 20% enrichment)30 min in buffermoseson2022b R14
same, cross-sectiondrug-rich boundary layer 94 ± 8 µm with crystals, F/F0 = 1.41 ± 0.10; gel layer below F/F0 = 0.93 ± 0.061 hmoseson2022b R15
same, confocalred layer 55 ± 6 µm at 5 min, 103 ± 9 µm at 30 minNile redmoseson2022b R16
50% BCL–PVPVA compact5–10 µm crust of crystals; red regions of 8 ± 2 µm at 5 min, a continuous layer of 1.9 ± 0.3 µm at 30 min, gone at 60 minSEM-EDX, confocalmoseson2022b R15, R17
10% BCL–PVPVA compactsurface smooth at 30 minSEMmoseson2022b R14
coated ritonavir–PVPVA tablet edgeconnected drug-rich regions along particle boundariesconfocal[benson2025 R14]
ritonavir–PVPVA 50%amorphous-ritonavir IR spectrum from 5 min; flaky at 15 minpH 6.8, 37 °C[indulkar2019a R16, R17]
ledipasvir–PVPVA 10%distinct top layer at 10 min; bulging, porous top by 30 minmicro-CT, dried[que2019 R13]
nilvadipine 15:85porous layer within 10 min, growing to 30 minmicro-CT[saboo2019 R13]
felodipine 30% in PVP, PVPVA, HPMCporous layer at the dissolving facemicro-CT, 10 min[saboo2020a R19]
felodipine 30% with 0, 2% TPGS; 5, 10% TPGSporous surface layer after 10 min; homogeneousmicro-CT, dried[saboo2021 R19]
indomethacin–PVPVA 15, 25%rough, porous surface after 5 min; smooth at 10%SEMsaboo2020 R19
Pitfalls and disagreements

The layers were measured on compacts, and the crystals were identified by shape and not by diffraction moseson2022b X20, X18. The polymer-depleted state of the 50% crust is inferred from the dye, and the authors read the crust as formed and dissolved repeatedly, which would make its slow release only "seemingly congruent" moseson2022b X16, C47. A drug-rich surface is not the only consideration: a TPGS dispersion released incongruently yet completely because little solid was left when nucleation began moseson2022b C50. The layer of the ITZ compacts is a "supposition" [bhardwaj2018 C52]; SEM and micro-CT samples were dried [indulkar2019a X11; que2019 X22].

← Concepts and formulas

concept · as used in Crystallization tendency

enthalpy of fusion

The heat absorbed when a crystal melts (ΔHfus, heat of fusion), measured by DSC; with Tm it gives the entropy of fusion and the Hoffman estimate of the crystallization driving force vaneerdenbrugh2010a E1, E2, and with the melting points of two polymorphs it decides monotropy by the heat-of-fusion rule lu2016 C50.

Findings

F6, F7, F13.

Related concepts

used by entropy of fusion, Hoffman equation and Burger-Ramberger rules; part of thermodynamic driving force for crystallization.

Formulas

Hoffman equation, entropy of fusion, Burger-Ramberger heat-of-fusion rule.

How it is measured or obtained

By DSC melting endotherms, in kJ/mol or J/g, and per volume with the crystal density baird2010 P1, P8; lu2016 R20.

Values in the library
materialvalue or classconditionspaper ids
ritonavir; haloperidol65.34; 54.26 kJ/molDSCbaird2010 M51, M17
caffeine20.81 kJ/molDSCbaird2010 M10
vemurafenib α; β53.8; 33.3 kJ/molDSClu2016 R20
nilutamide stable form; form I~98; around 66 J/gDSCtrasi2014b M1, R6
class I against class III≈+0.45 against ≈−0.63 SDper volumebaird2010 R22.b
nifedipine against felodipinehigher ΔHf and Tm, read as a higher intrinsic crystallization tendencyDSCmarsac2008a C35
Pitfalls and disagreements

One paper gives three different values for the α-vemurafenib heat of fusion, and its virtual transition temperature depends on the choice lu2016 X1; a melting endotherm overlapped by recrystallization can understate ΔHm lu2016 X12. Fusion properties did not correlate with crystallinity loss of drugs in PEO marsac2008b C34.

← Concepts and formulas

concept · as used in Crystallization tendency

entropy of fusion

The entropy gained on melting, \Delta S_{fus} = \frac{\Delta H_{fus}}{T_m} with Tm in kelvin baird2010 E26; vaneerdenbrugh2010a E1. It is read in two ways in the library: as part of the driving force for crystallization baird2010 C60 and, following Jackson, as a measure of how few melt molecules are correctly oriented to join the crystal trasi2014a C31. A related reading takes the entropy of crystallization as the probability that a molecule has the right conformation or orientation for nucleation marsac2006a C34.

Findings

F6, F13.

Related concepts

depends on enthalpy of fusion; part of thermodynamic driving force for crystallization; related to conformational flexibility.

Formulas

entropy of fusion, linear fit of normalized growth rate on melting entropy.

How it is measured or obtained

From DSC heat of fusion and melting temperature, per mole or, with the crystal density, per volume baird2010 P8; trasi2014a M1.

Values in the library
materialvalue or classconditionspaper ids
caffeine; haloperidol; ritonavir4.08, 12.8 and 16.4 × 10⁻² kJ mol⁻¹ K⁻¹DSCbaird2010 M10, M17, M51
felodipine; nifedipine74.4 and 89.6 J mol⁻¹ K⁻¹DSCmarsac2006a E11
nilutamide; cinnarizineΔSm/R ≈8.7; ≈12.5growth analysistrasi2014a R11.a, R11.i
class I against class III≈+0.41 against ≈−0.59 SDper volumebaird2010 R22.c
spin-coat class I against III≈+0.46 against ≈−0.33 SDper volumevaneerdenbrugh2010a R20.d
fast growers (group 1)≈−0.38 SDper moletrasi2014a R8.f
Pitfalls and disagreements

Fast melt crystallizers score high on ΔSfus per volume while fast growers score low on ΔSm per mole (synthesis: different bases and read-outs, see F6). For felodipine and nifedipine an entropy-dominated view would predict felodipine to crystallize more easily, which it does not; the enthalpy dominates there marsac2006a C35. Fusion properties did not correlate with crystallinity loss of drugs in PEO marsac2008b C34.

← Concepts and formulas

concept · as used in Crystallization tendency

entropy of mixing

The entropy gained when drug and polymer mix

For a drug with a polymer it is much smaller than with a small solvent, because the configurational entropy of the polymer is reduced by the connectivity of its repeat units, and it is nearly the same for any polymer grade marsac2006b C10, C11. For typical drugs (molecular weight 200–600) and polymers (10,000 and above) the contribution is "predicted to be relatively constant" and always favourable, so the enthalpy of mixing decides miscibility marsac2006b C12, C13; marsac2009 C26. In the ideal case the entropy of mixing of two components is ΔS = −R(X_u ln X_u + X_v ln X_v), printed without its minus sign in one paper [rumondor2009a E3, X7]. Where an entropy of mixing is not measured, readings of the activity coefficient and of χ as enthalpy rest on the assumption that it is close to ideal marsac2009 C11. For two polymers the entropy per lattice site falls with the chain length of both, from about −27 × 10⁻²³ J for a 783 Da maltodextrin with PVP K 12 to about zero for a 450 kDa dextran with PVP K 90, so the grade-independence found for a small drug does not hold between polymers [vaneerdenbrugh2012a R15, C34, C35].

Findings

F5, F9, F22.

Related concepts

part of free energy of mixing; includes combinatorial entropy of mixing; contrasts with enthalpy of mixing; related to Flory-Huggins theory.

Formulas

Flory-Huggins free energy of mixing, Gibbs free energy of mixing.

How it is measured or obtained

Not measured: it is computed from the Flory–Huggins expression with χ = 0, for a lattice whose site is one drug molecule marsac2006b P6, R1; for a small analogue liquid such as methyl pyrrolidone or 1-ethyl-2-pyrrolidone it is taken as ideal marsac2006b C46; marsac2009 C11.

Values in the library
materialvalue or classconditionspaper ids
nifedipine with methyl pyrrolidoneminimum near Φ ≈ 0.44χ = 0marsac2006b R1
nifedipine with PVP K12, K29/32, K90minimum near Φ ≈ 0.6; K29/32 and K90 virtually identical, K12 slightly lowerχ = 0marsac2006b R1
typical molecular weightsAPIs 200–600; polymers 10,000–1,000,000citedmarsac2009 R13
indomethacin, EP → PVP K12 → K29/32 → K90predicted solubility 0.396 → 0.144 → 0.134 → 0.134entropy term scaled with chain lengthmarsac2009 R9.e, C24
maltodextrin (783 Da) with PVP K 12 to K 90minimum about −27 to −21.5 × 10⁻²³ J per site25 °C, no enthalpy term[vaneerdenbrugh2012a R15]
6 kDa dextran with PVP K 12 to K 90−9.1 to −2.8 × 10⁻²³ Jsame[vaneerdenbrugh2012a R15]
450 kDa dextran with PVP K 90about 0same[vaneerdenbrugh2012a R15, C35]

Evidence table: entropy of mixing.

Pitfalls and disagreements

The generalization to all drugs and polymers rests on one simulated drug, with an unscaled axis, and on a PVP grade (K12, 2,500) below the stated polymer range marsac2006b X7, X8. The ideal-entropy assumption for the analogue liquid and for equal contacts in analogue and polymer was not tested marsac2009 C11, X17. In ternary systems with absorbed water the entropy still favours mixing, and demixing is expected only when the enthalpy of mixing is positive enough [rumondor2010c C27]. Two of the 20 plotted curves (PVP K 12 with 40 kDa and 450 kDa dextran) do not follow from the stated inputs [vaneerdenbrugh2012a X1].

← Concepts and formulas

concept · as used in Crystallization tendency

extended nucleation and growth classification

A risk classification that combines the DSC nucleation behaviour with the crystal growth rate: class IA (high nucleation and growth, overlapping zones, crystallizes on cooling), IB (high nucleation and growth, little overlap; good glass-forming ability, poor glass stability), II (rapid growth, low nucleation), III (ready nucleation, slow growth) and IV (slow nucleation and growth) trasi2014a C17. Risk for amorphous delivery is ranked I highest, then II, III intermediate, and IV "likely" the most robust trasi2014a C18.

Findings

F3, F15.

Related concepts

extends crystallization tendency classification; built on nucleation and crystal growth rate; related to glass stability.

Formulas

none.

How it is measured or obtained

From the cooling–reheating DSC class and the seeded growth rate of the neat drug trasi2014a C17, R1; nucleation itself is inferred, not measured trasi2014a C15.

Values in the library
materialvalue or classconditionspaper ids
carbamazepine, griseofulvinclass IADSC and growthtrasi2014a C17
acetaminophenclass IBDSC and growthtrasi2014a C17
nimesulide, nilutamide, celecoxibclass IIDSC and growthtrasi2014a C17
cinnarizine, droperidolclass IIIDSC and growthtrasi2014a C17
warfarin, miconazoleclass IVDSC and growthtrasi2014a C17
Pitfalls and disagreements

The labels reuse the numerals of the 2010 classes with different meanings, and some papers mix both schemes trasi2014a X10. The risk ranking assumes that growth-rate rankings above Tg carry over to storage temperatures trasi2014a C16, C35.

← Concepts and formulas

concept · as used in Crystallization tendency

fragility

How steeply molecular mobility or viscosity changes as a liquid approaches Tg: strong liquids follow Arrhenius behaviour, fragile ones depart from it baird2012 C37. It is expressed by the fragility index m (strong below 40, fragile above 75) or by the VTF strength parameter D (strong above 30, fragile below 10) baird2010 R30. Good glass formers were expected to be strong liquids baird2010 C3.

Findings

F5, F12.

Related concepts

measured by melt viscosity; depends on glass transition temperature; proposed predictor of glass-forming ability; related to reduced glass transition temperature.

Formulas

Vogel-Tammann-Fulcher equation.

How it is measured or obtained

m from the heating- and cooling-rate dependence of Tg (Moynihan) and D from m, or D from the VTF equation with T0 set to the Kauzmann temperature baird2010 P10, E14, E15, E18; D also by fitting the VTF equation to melt viscosity baird2012 P5, E4.

Values in the library
materialvalue or classconditionspaper ids
tolazamidem 18DSC rate dependencebaird2010 R18.k
itraconazolem 731DSC rate dependencebaird2010 R18.r
nifedipinem 33, D 36 from m, D 9 from VTF with TKDSCbaird2010 R18.i
28 compounds of classes I-B to IIID 4.3–8.7VTF fit of melt viscositybaird2012 R12
Pitfalls and disagreements

The category depends on the estimate used: aceclofenac, tolazamide and nifedipine are strong by m but fragile by the VTF-based D baird2010 X14. Fitted T0 values do not satisfy the paper's own T0–D–Tg relation baird2012 X5. Fragility did not separate good from poor glass formers baird2012 C43, and invoking it for outliers in growth data is untested trasi2014a X18.

← Concepts and formulas

concept · as used in Crystallization tendency

Gibbs-Duhem equation

The relation between the changes of chemical potential of the components of a mixture; integrated with the measured water activity against composition from a sorption isotherm, it gives the activity of the drug in the water-saturated amorphous phase, the term e^{-I(a_2)} of the amorphous solubility estimate (the Murdande method) almeidaesousa2015 E16, P7; ilevbare2013a P2. In the solution-crystallization papers the factor lowers the Hoffman estimate of the amorphous solubility, and I(a2) is the sum of a Henry's-law term from zero water to the limit of linearity and a numerical integral \sum \langle (n_1/n_2)(1/a_1)\rangle\,\Delta a_1 over the sorption isotherm up to unit water activity hsieh2012 E5, E6, P13; ilevbare2013c E6.

Findings

Amorphous solubility and LLPS: F5. Crystallization from solution: F2, F21.

Related concepts

yields the water sorption correction of amorphous solubility; input is water sorption; related to thermodynamic activity; corrects the Hoffman equation; used for the amorphous solubility advantage.

Formulas

Gibbs-Duhem equation, amorphous solubility advantage with water-sorption correction.

How it is measured or obtained

Numerical integration of moles of water per mole of drug against 1/a_1 over the sorption isotherm hsieh2012 E5. The papers of this chapter only name the method. The crystallization-from-solution papers measured the isotherm by dynamic vapour sorption of the amorphous solid at the test temperature (37 °C, 5–95% RH in 10% steps, three samples; 25 °C for danazol), with the water content extrapolated to water activity 1 [hsieh2012 P8, P13; jackson2014 P3, P4].

Values in the library

The resulting factors are listed under water sorption correction of amorphous solubility ilevbare2013a M1; almeidaesousa2015 M6. Values printed in the crystallization-from-solution papers:

materialvalue or classconditionspaper ids
danazol\exp[-I(a_2)] = 0.935DVS 5–95% RH, 25 °C[jackson2014 E1, R2]
five weak baseswater correction lowers the estimate for every base, for ketoconazole from 202 to 56.6 µg/mLDVS, 37 °Chsieh2012 R10.d
ritonavir, efavirenz, celecoxibI(a2) values not printed; theoretical amorphous solubilities 20.6, 19.8, 22.6 µg/mL37 °C[ilevbare2012b E8; ilevbare2013c E6, R2]
six dihydropyridinese^(−I(a2)) 0.74–0.91moisture sorption, TA Q5000raina2015b M1–M6
Pitfalls and disagreements

The expression is not printed in the 2015 and 2016 papers (named only) and the integral is not shown in any paper of 2013–2016 [almeidaesousa2015 E16; almeidaesousa2016 E4]. The sorption isotherms, the values of I(a2) and the preparation of the amorphous samples are not printed in the 2012 paper, so the estimates cannot be recomputed hsieh2012 X14. The amorphous solids for sorption (melt-quenched) and for dissolution (solvent-evaporated) differ in preparation ilevbare2013c X17. The sorption conditions and water contents are not printed raina2015b P3.

← Concepts and formulas

concept · as used in Crystallization tendency

glass-forming ability

In the library, glass-forming ability (GFA) is the ease with which a melt vitrifies on cooling instead of crystallizing; a compound that crystallizes during a controlled DSC cooling (class I) has a low GFA baird2010 C8; trasi2014a R14. It is kept apart from glass stability, the resistance of the glass to crystallization on reheating or storage baird2010 C4, C47; trasi2014a R14. The definition above comes from the crystallization-tendency papers, where GFA is measured on neat drugs from the melt. New aspect (ASD process): in the thermogravimetry study of 2020, indomethacin, posaconazole and bicalutamide were chosen as class III molecules of high glass-forming ability so that degradation could be tested in both the crystalline and the amorphous state; the class comes from the cooling and reheating classification of the library moseson2020 C5; baird2010 C8. Class III glass formers can still nucleate when held for an hour between Tg and Tm.

Findings

Crystallization tendency: F3, F9, F10, F11, F12, F15, F17. ASD process: F31. Physical stability: F17, F24, F34. Supersaturation from ASD dissolution: F7, F23.

Related concepts

contrasts with glass stability; measured by crystallization tendency classification and critical cooling rate; depends on nucleation temperature zone; related to melt viscosity and reduced glass transition temperature; see crystallization tendency; related to crystallization inhibition by polymers and nucleation and crystal growth; related to physical stability.

Formulas

Barandiaran-Colmenero critical cooling rate, Cabral critical cooling rate, critical cooling rate to avoid nucleation, reduced glass transition temperature.

How it is measured or obtained

By whether the melt crystallizes on DSC cooling at 20 °C/min, with liquid-N2 quenching to separate class I-A (still crystallizes) from I-B (vitrifies) baird2010 P1, P6, R3; by slow cooling at 1 °C/min baird2010 R7; by the critical cooling rate from rapid heat–cool DSC baird2013 R5, R9; and, as a proxy, by the melt viscosity at Tm baird2012 C31. A drug's GFA is also reported from DSC cooling in later formulation papers trasi2020 R5; duong2022a R3. In the 2020 thermogravimetry study the class was cited, not measured; what was measured is nucleation of the neat undercooled melt during isothermal holds below Tm: posaconazole (130–160 °C, 1 h) and bicalutamide (130–180 °C, 1 h) melted on reheating, indomethacin (130–150 °C, 4 h) showed only a glass transition moseson2020 P7, R7. When the melt route is excluded by degradation, by attempting amorphization by spin coating, rotary evaporation and cryomilling wegiel2013b P2, R1. In the study of commercial tacrolimus capsules: DSC of crystalline tacrolimus monohydrate, with a first heating (broad endotherm between 60 and 120 °C, then melting of the anhydrate), cooling of the melt to an amorphous solid, and reheating through the glass transition without crystallization; the high Tg and the absence of recrystallization on reheating "suggest" good glass-forming ability and stability trasi2016 R3, M1, C13.

Values in the library
materialvalue or classconditionspaper ids
51 drug-like molecules23 crystallize on cooling (low GFA)DSC, 20 °C/minbaird2010 R1
23 class I molecules8 vitrify on liquid-N2 quench (I-B)melt plunged into liquid N2baird2010 R3
antipyrin, chlorpropamide, flufenamic acid, tolbutamideglass already at 25 °C/minrapid heat–cool DSCbaird2013 R5
8 class I-A moleculescrystallize even at 800–1100 °C/minrapid heat–cool DSCbaird2013 R1, R3
acetaminophenforms a glass even at slow coolingcited DSC datatrasi2012a R1
nilutamidegood glass former; attributed to low nucleationDSC at 15 °C/mintrasi2014b R2, C9
vemurafenib"relatively poor glass former", about 25 °C supercoolingDSC at 10 °C/minlu2016 R1, C5
tacrolimusgood GFA and stability; melting onset 121 °C, Tg onset 75 °C, no crystallization on reheatingDSC heat–cool–reheattrasi2016 R3, C13, M1
lumefantrine; artemetherclass 3 (Tg ≈18 °C, no crystallization on reheating); class 1 (crystallized on cooling from about 60 °C)DSC heat–cool–heattrasi2020 R5, R6, C7
acetaminophenglass because nucleation (50–60 °C) and growth barely overlapDSC cool–reheat, hot stagetrasi2012a C39, R12.a, R8.a
resveratrolnot made amorphous by spin coating from four solvents, rotary evaporation or 6 h of cryomillingneat drugwegiel2013b R1, C6
curcuminamorphous by spin coating, not in larger amounts by rotary evaporation; cryomilling gave a larger amorphous massneat drugwegiel2014 C9
celecoxibeasily made amorphous by cooling the melthot stagemosqueragiraldo2014 R15
delamanidno glass at up to 50 °C/min; class I; a glass quenched in liquid nitrogen recrystallized just above its Tg of 43.3 °CDSC; liquid-N2 quenchduong2022a R3, C44
bicalutamidehigh GFA but rapid crystallization above Tgcitedmoseson2022b R26
posaconazole, indomethacin, bicalutamideclass III (high glass-forming ability)citedmoseson2020 C5
posaconazole, bicalutamide neat meltsnucleated within 1 h between Tg and TmDSC holdsmoseson2020 R7
Pitfalls and disagreements

A high GFA says nothing about stability on reheating: class II and some class III drugs form glasses easily yet crystallize on reheating or grow fast once nucleated baird2013 C24; trasi2014a C14. The judgement depends on the cooling rate and on the unknown liquid-N2 quench rate baird2010 C17, C21. Standard predictors (Tg/Tm, glass-stability parameters, fragility) did not rank GFA of organic molecules baird2010 C38; baird2012 C43. The amount of polymer needed for a stable dispersion is expected to depend on GFA and glass stability trasi2012a C4. A high glass-forming ability also says nothing about stability of the glass on holding near Tm; the dispersion, not the neat glass, was used for isothermal degradation tests for that reason moseson2020 C17, C55. Easy glass formation said nothing about growth once nucleated: celecoxib forms a glass easily yet grows fast mosqueragiraldo2014 R15, R16. Good glass-forming ability of lumefantrine, read as making it "a good candidate" for a stable dispersion, did not keep its PVPVA dispersion amorphous beyond 1 week at 40 °C/75% RH trasi2020 C27, R3. The tacrolimus result rests on one cooling run, at 15 °C/min with reheating at 10 °C/min trasi2016 P1; the class is inferred from it, not measured by the cooling-rate criteria of the crystallization-tendency papers trasi2016 C13. Rotary evaporation was too slow to trap delamanid, a poor glass former, above about 10–15% drug, whereas spray drying gave an amorphous dispersion at 25% drug [duong2022a C45; nguyen2023c R5].

← Concepts and formulas

concept · as used in Crystallization tendency

glass-stability parameters

Combinations of Tg, the onset and peak of crystallization on reheating and Tm (KLL, KH, KW, KSP, K1, K2 and the reduced temperature Tred), taken from inorganic glass science; a larger gap between Tg and crystallization onset "generally" means a better glass former baird2010 E5, E11, C2.

Findings

F12.

Related concepts

measures glass stability; related to reduced glass transition temperature and cold crystallization.

Formulas

none with a page; the parameters are listed in the chapter formula table.

How it is measured or obtained

From DSC reheating of glasses made at a given cooling rate baird2010 P11; for inorganic glasses the parameters rose as the measured critical cooling rate fell baird2010 R28.

Values in the library
materialvalue or classconditionspaper ids
acetaminophen; celecoxibTred 0.33; 0.72after 20 °C/min coolingbaird2010 R16.a, R16.c
acetaminophen; celecoxibKH 0.474; 2.532DSCbaird2010 R17.d, R17.f
celecoxiblowest KSP (1.464) and K2 (1.118) of class IIDSCbaird2010 R17.f
tolazamidehighest KSP (5.263) and K2 (3.452) of class IIDSCbaird2010 R17.n
cinnarizineTred 0.36 after 1 °C/min and 0.56 after 20 °C/min coolingDSCbaird2010 R16.d
Pitfalls and disagreements

The parameters ranked the wrong class II molecules as the best glass formers baird2010 C28, and the ranking depends on which parameter is used baird2010 X15. They cannot be reproduced from the printed temperatures, and one pair of values is misprinted baird2010 X8, X1. For five pharmaceutical glasses a higher Tred had gone with faster crystallization (cited) baird2010 R28.

← Concepts and formulas

concept · as used in Crystallization tendency

glass stability

The resistance of a glass, or of the liquid formed when it is reheated above Tg, to crystallization; the authors of the 2010 classification call the term "somewhat of a misnomer" because it is usually judged by reheating rather than by the glass itself baird2010 C4. In the classification, glass stability means no crystallization on reheating trasi2014a R14.

Findings

F9, F15, F16, F17.

Related concepts

contrasts with glass-forming ability; measured by glass-stability parameters and critical heating rate; depends on quenched-in nuclei; part of physical stability.

Formulas

critical cooling rate to avoid nucleation.

How it is measured or obtained

By DSC reheating after cooling (class II against III) baird2010 R1; by glass-stability parameters from crystallization onsets on reheating baird2010 E5; by critical heating rates baird2013 R9; by storage of glasses below Tg baird2010 P7.

Values in the library
materialvalue or classconditionspaper ids
class IImuch lower glass stability than class IIIDSC reheating and storagebaird2010 C47
class IIIhigh glass-forming ability and high stability on reheatingDSCbaird2010 C34
indoprofen glassfully crystalline after 1 day30 °C, 0% RHbaird2010 R23
class II glassesRcrit,heat 12.5 to above 1910 °C/minrapid heat–cool DSCbaird2013 R9
acetaminophenquenched-in nuclei may matter for stability over longer timesargumenttrasi2012a C41
Pitfalls and disagreements

The storage evidence for a link between glass-forming ability and glass stability rests on one or two glasses per class baird2010 X16. Glass-stability parameters ranked class II molecules inconsistently baird2010 X15.

← Concepts and formulas

concept · as used in Crystallization tendency

glass transition temperature

The temperature that separates the glass from the supercooled liquid lu2016 C2; it is a kinetic transition where the system falls out of equilibrium trasi2014b C2. Tg is called "probably" one of the most important properties of an amorphous solid, because storage temperature relative to Tg affects stability towler2008 C20. This draft is written from the crystallization-tendency chapter; the evidence table covers far more papers that later chapters will add. New aspect (PEG dispersions): the Tg of the drug relative to the solidification temperature as a possible determinant of whether an amorphous drug is excluded from the crystalline carrier. In polymer blends the additive's mobility, through the blend Tg, is one of three factors that decide exclusion zhu2011 C2. In the drug/PEG study a drug Tg above the 25 °C solidification temperature (nilutamide about 33 °C, ketoconazole 44 °C) did not change the pattern zhu2011 C28, C31. For water in PEO, Tg describes plasticization (water −136 °C, PEO about −67 °C) marsac2008b M12, M7, C26.

Findings

Crystallization tendency: F4, F12, F20. PEG dispersions: F6, F11.

Related concepts

used in reduced glass transition temperature and Tm − Tg temperature gap; bounds the supercooled liquid; related to fragility, amorphous salt, plasticization and interlamellar incorporation.

Formulas

reduced glass transition temperature, Vogel-Tammann-Fulcher equation.

How it is measured or obtained

By DSC on reheating a melt-quenched or freeze-dried sample, as onset or midpoint baird2010 P1; trasi2014b M1; towler2008 P6; for fast crystallizers that give no Tg, estimated from a Tg/Tm ratio or a Trg–molecular-weight fit vaneerdenbrugh2010a E7; baird2010 E27. In the PEG papers the Tg values of the drugs were cited from the crystallization-tendency classification, or measured by DSC (acetaminophen) or estimated with a Tg/Tm ratio (naproxen) zhu2011 M5, M6; marsac2008b M1, M3, E5.

Values in the library
materialvalue or classconditionspaper ids
propranolol base; hydrochloride−9.3; 35.9 °CDSCtowler2008 R4
nicardipine base; hydrochloride3.2; 81.0 °CDSCtowler2008 R5
nilutamide32 °C (onset)DSCtrasi2014b M1
vemurafenib108 °CDSC at 10 °C/minlu2016 M1
felodipine; nifedipine; indomethacin45 °C eachDSC midpointbaird2010 M37, M32, M40
lumefantrineabout 18 °C; 19.7 °C (onset)DSCtrasi2020 R5; hiew2022a R3
five drugs with 10 mol% PVPraised by about 2–3 °CDSCtrasi2014a R7
nilutamide; ketoconazoleabout 33 °C; 44 °Ccitedzhu2011 M5, M6
benzocaine−31 °Ccitedzhu2012 M3
acetaminophen; ketoprofen; naproxen; ibuprofen; felodipine; ketoconazole22.1; −3.0; 24 (estimated); −43.6; 43.1; 44.4 °CDSC and citedmarsac2008b M1, M2, M3, M4, M5, M6
amorphous PEO; waterabout −67 °C (reported −67 to −55 °C); −136 °Ccitedmarsac2008b M7, M12
Pitfalls and disagreements

Whether a Tg is onset or midpoint is not always stated towler2008 X7; Tg of 16 class I molecules is an estimate baird2010 X17. Crystallization tendency cannot be predicted from relaxation times and Tg alone marsac2010 C29, and a high Tg did not prevent a propranolol salt from crystallizing towler2008 R4.a, R12. In the solvent screen the Tg contribution separates the classes partly because the storage criterion selects high-Tg compounds vaneerdenbrugh2010a C41. The Tg conclusion in the drug/PEG study rests on two drugs whose Tg lie 8 and 19 °C above 25 °C and that also differ in interaction and crystallization tendency zhu2011 X13. The Tg of haloperidol is given as below 25 °C in the card of Zhu and co-workers and as 33 °C (calculated from the molecular weight) in the classification paper zhu2011 M1; baird2010 M17. The naproxen Tg of 24 °C does not follow from the Tg/Tm rule it cites: the Table 1 melting point gives 26.7 °C marsac2008b E5, X4. (synthesis: papers also give different Tg for the same drug, nilutamide 32 °C (onset) against about 33 °C, and felodipine 45 °C (midpoint) against 43.1 °C trasi2014b M1; zhu2011 M5; baird2010 M37; marsac2008b M5.)

← Concepts and formulas

concept · as used in Crystallization tendency

heterogeneous nucleation

Nucleation at an interface (a foreign particle, a container wall, a second phase), which lowers the nucleation barrier, so nucleation there proceeds more readily wilson2021 C44. New aspect (PEG dispersions): a crystalline carrier as the nucleating surface. Zhu and co-workers conclude that crystalline PEG acts as a favourable heterogeneous nucleation site for the slowly crystallizing ibuprofen and fenofibrate, which explains why the drugs began to crystallize in the dispersion (after about 100 and 120–180 min) earlier than most neat samples, despite a smaller thermodynamic driving force ("it appears") zhu2012 C4, C34, R23.a. New aspect (Crystallization from solution): In solution, impurities in large volumes at high supersaturation may induce heterogeneous nucleation, which the authors give as a reason for the small scatter (below 15%) of induction times ("it is reasonable to expect") ilevbare2013c R17, C16. For danazol the interface of the drug-rich droplets is a candidate site: crystals could form from the drug-rich or the drug-lean phase, the change to a two-phase solution can alter nucleation, and polymer at the droplet–water interface is "likely" to disrupt nucleation there [jackson2014 C45, C53]. Micelles of dihydroxy bile salts "may" offer interfaces for heterogeneous nucleation, and the surfaces of drug-rich telaprevir particles are expected to enable it; adsorbed bile salt is expected to modify it lu2017 C34, C36, C39. New aspect (Supersaturation from ASD dissolution): the ASD dissolution papers invoke it three times: polymer adsorbed on parent γ nifedipine crystals could reduce heterogeneous nucleation of α on them, and many small solution-grown crystals offer more surface for it raina2014b C31, C36; the interfaces of a drug-rich phase "may" act as heterogeneous nucleation sites [trasi2015b C31]; surfactants promoted ritonavir crystallization only when the ritonavir–water interface of droplets was present [ilevbare2013d C43].

Findings

Crystallization tendency: F10, F17. PEG dispersions: F9. Crystallization from solution: F14, F15, F25, F32, F34, F37, F38. Supersaturation from ASD dissolution: F3, F11, F13.

Related concepts

part of nucleation; contrasts with classical nucleation theory (homogeneous form); related to surface crystallization, stochastic nucleation, nucleation induction time and semicrystalline solid dispersion; related to secondary nucleation and polymer adsorption to drug-rich droplets; related to drug-rich nanodroplets and surfactant-induced crystallization.

Formulas

classical nucleation theory.

How it is measured or obtained

Rarely measured directly; inferred from scatter of crystallization temperatures in DSC pans baird2010 C19, from crystals forming at a seeded interface zhu2012 R15, R19, from the effect of shear in a rheometer baird2012 C8 or from the change of slope of induction time against supersaturation duong2022b C28. In the seeding test a small piece of crystalline PEG is placed in the supercooled neat drug at 25 °C and observed by polarized light microscopy for where the first crystals form; crystals formed at the PEG edge within minutes and none in the bulk zhu2012 P7, R15, R19. In the crystallization-from-solution papers it is not measured; the site of nucleation (droplet, interface or bulk) was not determined [jackson2014 X15]. Seeds added on purpose test the related seeded growth (F5, F20) [alonzo2012 P4]. In the ASD dissolution papers it is not measured either, but argued from conversion kinetics and induction times [raina2014b C32; ilevbare2013d R16].

Values in the library
materialvalue or classconditionspaper ids
ibuprofen; fenofibratecrystals form at a crystalline PEG interface within minutes, none in the bulk25 °Czhu2012 R15, R19, C4
neat ibuprofen; fenofibrate0.20 and 0.34 of 50 samples crystallized after 3 weeks25 °C, no carrierzhu2012 R2, R3
ibuprofen; fenofibrate in 20/80 PEG 3350onset about 100 min; 120–180 min25 °Czhu2012 R23.a
class I-A meltscrystallization temperatures higher in the rheometer than in DSC, ascribed to the upper plateshear 30 s⁻¹baird2012 R5, C8
class I moleculesscatter of crystallization temperatures ascribed to stochastic and heterogeneous nucleationDSC pansbaird2010 C19; baird2013 C18
felodipine filmsnucleation "most likely heterogeneous"spin-coated filmskonno2006 C33
posaconazolehomogeneous at high supersaturation, heterogeneous at low; shallow slope of log t against 1/log²S below S about 12 read as heterogeneous ("has been attributed")PBS pH 6.5, 37 °Cduong2022b R10, C28
enzalutamide with drug-rich dropletsdroplets enhance crystallization; droplet/water interface "likely" a favourable site; induction times fall on crossing 42 µg/mLpH 6.5, 2.5–100 µg/mL polymerwilson2021 C45, C46, R3
bicalutamidefractures of mechanically activated samples favour heterogeneous nucleationdispersionsmoseson2022b C45
induction-time replicates, 50–80 mL, S 2.4–15.4scatter below 15%triplicateilevbare2013c R17
danazol, HPMC-ASinduction time 516 → 277 min as drug concentration rises from 10 to 20 µg/mL25 °C[jackson2014 R17]
telaprevir δ 28.8, buffercrystallizes immediately with a drug-rich phase presentpH 6.5, 37 °Clu2017 R4
atazanavir ASD particlesdissolving ASD did not seed nucleation (about 300 against 350 min)PVPVA + SDS, pH 6.8[indulkar2019b R5, C25]
ritonavir with SDS; Tween 80, below and above LLPSabout 262 against 34.7 min; about 289 against 23.6 min37 °C[ilevbare2013d R16, R12]
nifedipine dispersions against glassγ → α complete in ≈5–40 min against ≈40–145 min with dissolved polymer25 °Craina2014b R12, R10
Pitfalls and disagreements

Heterogeneous nucleation by a carrier breaks the link between the crystallization of the pure drug and of the drug in the dispersion zhu2012 C41. Homogeneous classical theory is applied to systems that are most likely heterogeneous konno2006 C33; marsac2006a C32. The PEG seeding was done in pure supercooled drug, not in the dispersion, and enrichment of the drug in the remaining non-crystalline fraction after PEG crystallizes is an alternative that the experiments cannot separate zhu2012 X13. Whether other slowly crystallizing drugs (nilutamide, ketoconazole) are nucleated by crystalline PEG was not tested; in the same PEG at the same ratio neither was fully crystalline at 24 h zhu2011 R18.e, R18.f. Nucleation by polymers in solution has been reported, outside the library, and most reports of drugs in polymers find retardation of crystallization zhu2012 C35, C36. The attribution to nucleation at the droplet interface is an inference; solutions at 10 µg/mL with HPMC or HPMC-AS are already above their measured onsets [jackson2014 X3, X15]. Impurity-induced nucleation was expected, not tested ilevbare2013c C16. Neither micelles as nucleation sites nor the nucleation site in the two-phase solutions were observed lu2017 C34, C40. Neither the enzalutamide nor the posaconazole paper located the nucleation site; the homogeneous–heterogeneous assignment rests on a slope change wilson2021 C45; duong2022b X11. None of the three proposals of the ASD dissolution papers was tested directly [raina2014b C36; trasi2015b C31; ilevbare2013d C43].

← Concepts and formulas

concept · as used in Crystallization tendency

Hoffman equation

An approximation for the free-energy difference between the supercooled liquid and the crystal at a temperature T below Tm, needing only Tm and the heat of fusion vaneerdenbrugh2010a E2; lu2016 E1. It assumes that the enthalpies of crystal and supercooled liquid vary linearly with temperature, that is a constant heat-capacity difference, and applies above Tg marsac2006b C37; ilevbare2013a C58. In this chapter it supplies the crystallization driving force; most other library papers use it to estimate the amorphous solubility advantage. New aspect (solution methods): in the solution-methods papers the Hoffman equation supplies the free-energy advantage of the amorphous form in the amorphous solubility estimate, printed with a positive sign and \Delta T = T_m - T: \Delta G = \frac{\Delta H_f\,\Delta T}{T_m}\cdot\frac{T}{T_m} raina2015a E2. New aspect (C. Amorphous solubility and LLPS): those papers print it as the free-energy term of Eq. 1, \Delta G = \Delta H_f\,(T_m - T)\,T/T_m^{2}, and list its assumptions: a constant, temperature-independent ΔCp, small undercooling and Tg close to T∞; it also does not address the crystal-to-glass conversion [ilevbare2013a E2, C58; almeidaesousa2015 E2, C8; mosqueragiraldo2015 E2; almeidaesousa2016 E2]. For atazanavir Eq. 2 is printed with DSC Tm 208 °C and ΔHf 90.1 J/g, and the paper's own ratio of 62 follows from them when 0.22 is read as exp[−I(a)], although Table I heads that column I(a) (synthesis) [indulkar2017a E2, E3, X1; indulkar2015 M2]. New aspect (Bioperformance): the equation estimates the amorphous-to-crystalline solubility ratio \sigma_a/\sigma_c = e^{\Delta G/RT} and is combined with a water-sorption correction [raina2014a E3, E5; raina2015c E1, E2]. New aspect (Crystallization from solution): in the solution-crystallization papers \exp(\Delta G_c/RT) is the activity ratio taken as the solubility ratio before a water-sorption correction, evaluated at 25 or 37 °C, and the ratio times the measured crystalline solubility is compared with plateaus after pH shifts and with onsets of phase separation (F2) [hsieh2012 E3, E4, R10; hsieh2014 E2, E3; jackson2014 E2, R5]. New aspect (Supersaturation from ASD dissolution): the ASD dissolution papers use it to estimate the amorphous solubility advantage of felodipine and indomethacin; the felodipine inputs of 2008 come from earlier work, which found the equation estimated felodipine's free-energy difference well (cited) [konno2008a E2, R20].

Findings

Crystallization tendency: F6, F7, F13. Solution methods: F5. C. Amorphous solubility and LLPS: F5, F7. Bioperformance: F1, F23. Crystallization from solution: F2, F16, F21. Supersaturation from ASD dissolution: F5, F7.

Related concepts

measures thermodynamic driving force for crystallization; depends on enthalpy of fusion and degree of supercooling; related to entropy of fusion, amorphous-crystalline free energy difference and amorphous solubility; compared with the heat-capacity route in amorphous-crystalline free energy difference; accuracy set by degree of supercooling. Related to amorphous solubility advantage; corrected by the Gibbs-Duhem equation route.

Formulas

Hoffman equation, amorphous solubility advantage from free energy difference.

How it is measured or obtained

Calculated from DSC Tm and ΔHfus at the temperature of interest, per mole, per gram or, with the crystal density, per volume baird2010 E25; vaneerdenbrugh2010a P11. Its agreement with MDSC-based free energies was checked for felodipine and nifedipine marsac2006a C14. In solution work it enters \exp\left(\frac{\Delta G}{RT}\right) as the amorphous-to-crystalline solubility ratio alonzo2010 R3; duong2022a R27. ΔHf and Tm were measured by DSC at 10 °C/min in pinhole aluminum pans; the values are not printed raina2015a P4, E2. With a heat-capacity term it becomes \frac{\Delta G_{a\to c}}{RT} = \frac{\Delta H_f}{RT}\left(1 - \frac{T}{T_m}\right) + \frac{1}{RT}\int_{T_m}^{T}\Delta C_p\,dT - \frac{1}{R}\int_{T_m}^{T}\frac{\Delta C_p}{T}\,dT mosqueragiraldo2015 E3. In the amorphous solubility papers ΔHf and Tm are DSC values at 10 °C/min (5 °C/min in one paper), evaluated at 25 or 37 °C [almeidaesousa2015 P5; indulkar2015 P6, P5; almeidaesousa2016 P9]. In the Bioperformance papers ΔHf and Tm come from DSC melting onset and enthalpy, and the water sorption from gravimetric moisture sorption raina2014a P3, P4. The DSC method is not given in the 2014 pH-shift paper and the heats of fusion are not printed in the 2012 paper hsieh2012 P13, X14; hsieh2014 X11. In the ASD dissolution papers: from the DSC enthalpy of fusion and melting temperature (felodipine 30.12 kJ/mol, 143.41 °C; indomethacin 32.48 kJ/mol, 153.4 °C) and the crystalline solubility alonzo2010 P10, P12, M1, M2; inputs not printed in the 2008 and 2011 papers [konno2008a E2; alonzo2011 E3].

Values in the library
materialvalue or classconditionspaper ids
felodipine; nifedipine6.2 and 8.8 kJ/mol25 °Cmarsac2006a E3, R4
34 drugsΔGv −8.6 (lidocaine) to −54.7 J/cm³ (acetaminophen)298 Kbaird2012 M5, M12
vemurafenib α against βα lower in free energy up to a virtual transition at 319 °CTm and ΔHm of both formslu2016 R22, E1
felodipineestimated amorphous solubility about 9 µg/mL37 °Calonzo2010 R3
delamanidsolubility ratio 31.8 calculated against 42.3 measured37 °Cduong2022a R27
ritonavir, efavirenz, celecoxib≈10.2, ≈2.70, ≈7.74 kJ/mol (extractor calculation)37 °Cilevbare2013c E7
six dihydropyridinesused for the yardstick of F5; no ΔG value printed25 and 37 °Craina2015a E2, R11
diverse drugsCp method closer to the measured LLPS onsets than the Hoffman equation25 °Calmeidaesousa2015 R11, C43
ritonavir10.1 kJ/mol (extractor recomputation from Table 2 inputs)37 °Cilevbare2013a E2
ibuprofen to griseofulvin3.24, 4.84, 5.69, 8.51, 10.33, 9.69 kJ/mol25 °Calmeidaesousa2015 R11
telaprevir13.31 kJ/mol (recomputed); amorphous solubility 245 ± 5 µg/mL, 2.7 times the onset37 °Cmosqueragiraldo2015 E1, R13, R17
clotrimazole6.26 kJ/mol with a molar mass not printed in the paper (extractor)310 Kindulkar2015 E3
CRH15.68 kJ/mol; estimate 11.2 µg/mL at pH 4.625 °C[almeidaesousa2016 M1, R2]
atazanavir14.55 kJ/mol (recomputed from Table I); ratio about 62, as printed, with 0.22 read as exp[−I(a)]; about 228 read as I(a)310 K[indulkar2017a E2, X1; indulkar2015 M2]
felodipineHoffman ratio 10.92 (25 °C), 8.64 (37 °C); water factor 0.74, 0.83Tm 147 °C, ΔHf 28.67 kJ/molraina2014a R7
nifedipineprinted ratio 8.78; recomputed 14.45Tm 173 °C, ΔHf 32.5 kJ/molraina2014a R7, X3
atazanavirratio 62, predicted 68 µg/mL310 K[indulkar2017a R5]
indomethacin17–25 µg/mL (25 °C), about 37 µg/mL (37 °C)Hoffman estimatealonzo2010 R8
enzalutamideprinted estimate about 39 µg/mL (13-fold); the printed inputs give 27.737 °C, Tm 470 K[wilson2018 E1, R3, X1]
ritonavirΔHf 60.4 kJ/mol, Tm 121 °C, 37 °C: ΔG = 10.1 kJ/mol and exp(ΔG/RT) = 50.8 (extractor calculation); calculated amorphous solubility 20.6 µg/mL with the water term37 °C[ilevbare2012b E8, E9, R1]
glyburide; warfarinabout 60-fold and 48-fold (59.9 and 47.7 recomputed) from Tm 175.6 °C, ΔHf 45.5 kJ/mol and Tm 169.2 °C, 43.6 kJ/mol25 °C (assumed)hsieh2014 R16, E3, M3, M4
danazolTm 225.4 °C, ΔHf 28.9 kJ/mol: ΔG = 6.95 kJ/mol; ratio with the 0.935 water factor 15.4 recomputed against 19.6 printed25 °C[jackson2014 E1, E2, X1]
five weak basesratios 22.2, 11.9, 20.4, 54.6, 7.0 without water correction (carvedilol, clotrimazole, clozapine, ketoconazole, loratadine)37 °Chsieh2012 R10
felodipine (six-dihydropyridine study)e^(ΔG/RT) recomputed 10.84 at 298.15 K from 74.6 J/g and 147 °CDSC data of the paperraina2015b E1, R1.b
posaconazoleTm 169.1 °C, ΔHf 76.95 J/g; printed amorphous/crystalline ratio 37.9 with water factor 0.33, estimate 10.22 against 9.08 µg/mL measured310 K; the extractor obtains ΔG = 16.13 J/g and a ratio of about 26.5duong2022b R7, E3, X2
felodipineratio about 10; about 9.5 µg/mL37 °C[konno2008a R17]
felodipineΔG 5729 J/mol, ratio 9.22 (recomputed) against "around 9"37 °Calonzo2010 E1
felodipineabout 9.1 µg/mL (37 °C); 5–6 µg/mL (25 °C)pH 6.8[alonzo2011 M1]
Pitfalls and disagreements

The approximation "can lack accuracy": it assumes a constant ΔCp and small undercooling almeidaesousa2015 C8, C34, and was judged inappropriate for telaprevir, far below its melting point mosqueragiraldo2015 C24. It ignores ionization and water sorption raina2014a C10. Printed values are sometimes not reproducible from the printed inputs baird2012 X2; raina2014a X3. Sign conventions differ: some papers plot crystal minus liquid lu2016 X17 or print a leading minus sign duong2022a E2. In the solution-methods papers the Hoffman-based estimate was above the measured danazol onset (16 against 13 µg/mL) and cannot follow polymer effects [jackson2014 R5]. In the amorphous solubility papers, for telaprevir (Tm 238 °C, 37 °C test) the authors judge it inappropriate for large undercooling mosqueragiraldo2015 C24, M1. In the 2013 paper four of eight predictions cannot be reproduced from the printed Table 2 inputs ilevbare2013a X1. Eq. 2 of the weak-base paper is printed with a sign that does not give its stated ratios indulkar2015 X1. Eq. 3 of the atazanavir paper is printed with an inverted sign [indulkar2017a X1]. The nifedipine ratio of raina2014a does not follow from the printed inputs raina2014a X3. The enzalutamide estimate cannot be reproduced from its printed inputs, and the stated S of "approximately 13" matches the estimate rather than the measured 42/2.9 = 14.5 [wilson2018 X1, X2]. The danazol ratio cannot be reproduced from the printed inputs, and the weak-base estimates cannot be recomputed from the 2012 paper [jackson2014 X1; hsieh2012 X14]. The equation neglects water mixed into the amorphous phase, which can lower the amorphous solubility by up to a factor of 2 hsieh2014 R17. Its use for warfarin assumed 25 °C because the temperature is not printed hsieh2014 E2. For posaconazole the printed ratio of 37.9 does not follow from the printed melting data, and the exponent of the printed amorphous-solubility equation has the wrong sign duong2022b X2. The indomethacin estimate at 25 °C does not reproduce from the printed inputs alonzo2010 X2. The estimate becomes increasingly approximate far below Tg, and a configurational heat-capacity estimate is more rigorous (cited) [alonzo2011 C5].

← Concepts and formulas

concept · as used in Crystallization tendency

hydrogen bonding

In the papers of this chapter, a bond between a donor group of the drug (the carboxylic acid OH of indomethacin, the NH of felodipine and nimesulide, hydroxyl groups of sucrose) and an acceptor group of the polymer (the lactam carbonyl of PVP, the acetate groups of PVAc and PVP/VA, the carboxyl of PAA, the groups of HPMCAS), or between molecules of the same kind (acid dimers of the drug, self-association of PAA) [taylor1997 C25, C35; kestur2010a C18; rumondor2009a C45]. A lower NH wavenumber means a stronger bond kestur2010a C18. PVP carries a better acceptor than the drug's own groups, so its bonds replace weaker drug–drug bonds marsac2009 C39; the more acidic carboxyl of indomethacin is expected to bond more strongly than the NH of felodipine or nifedipine or the OH of sucrose marsac2009 C42. A drug donor can also be bound within its own molecule (see intramolecular hydrogen bonding), which leaves little to bond to a polymer kestur2011 C27. Later papers add polyphenol OH groups bonding to the PVP carbonyl (a lower OH wavenumber and a larger carbonyl shift mean a stronger bond: OH band at 3138 and 3135 cm⁻¹ with PVP against 3329–3399 cm⁻¹ with the other polymers) wegiel2013a C8, C9, C11, R2, R4; dextran hydroxyls bonding to the PVP carbonyl in a polymer–polymer pair, where dextran–dextran bonds are assumed to be stronger or more numerous [vaneerdenbrugh2012a C10, C36]; and a donor and acceptor classification of the groups of 15 drugs and 7 polymers by bonding atom, acidity and the pKBHX scale [vaneerdenbrugh2011a C13, C14, C19]. Lumefantrine did not hydrogen-bond to PVPVA in dried films hiew2022a C20, R17.

Findings

F1, F2, F3, F4, F12, F13, F14, F17, F20, F22, F24, F25.

Related concepts

part of drug-polymer interactions; includes intramolecular hydrogen bonding and self-association; contrasts with dipole-dipole interactions; related to hydrogen-bond basicity, pKa and binding constant; contrasts with halogen bonding and acid-base interaction; classified by Etter's hydrogen-bond rules.

Formulas

fraction of hydrogen-bonded carbonyl groups from band areas.

How it is measured or obtained

By IR and Raman. Cyclic acid dimers are found by the mutual exclusion rule: an acid band present in the IR and absent in the Raman taylor1997 C14. Drug–polymer bonds are read from shifts of the drug NH or acid C=O band and of the polymer carbonyl band, in spin-coated or solvent-cast films at one donor per polymer carbonyl or at 20–50 wt% polymer, always against the amorphous drug and a physical mix, not against the crystal marsac2009 P4; kestur2010a P6, P7; taylor1997 C7, C24. A dilution series in a non-polar solvent tests for intramolecular bonds kestur2011 P9. In dispersions, a second principal component of the IR spectra beyond a calculated physical mixture is read as "presumably" hydrogen bonding [rumondor2009a C20]. DFT gives the geometry of single molecules kestur2011 P11. Further methods: the fraction of PVP carbonyls bonded, from a free and a maximally bonded band fitted to the carbonyl region (fraction of hydrogen-bonded carbonyl groups from band areas) [vaneerdenbrugh2012a P5, E1]; the polyphenol OH band and the polymer C=O shift at 30% polyphenol wegiel2013a P4; crystal-structure statistics of competition between groups [vaneerdenbrugh2011a P5, R14]; DFT for intramolecular bonds [vaneerdenbrugh2011a P6].

Values in the library
materialvalue or classconditionspaper ids
indomethacin–PVP-K90PVP C=O at 1636 against 1679 cm⁻¹; drug acid C=O at 1726 cm⁻¹5–70 wt% PVP, solvent-evaporatedtaylor1997 R5.l, R11, R12
amorphous indomethacinacid dimer at 1710 cm⁻¹ (IR) with no Raman bandquench-cooled melttaylor1997 R5.h, R6
indomethacin, felodipine, sucrose, ketoconazole with PVP K12PVP C=O shift 45, 25, 25 and 0 cm⁻¹one donor per carbonyl, spin-coatedmarsac2009 R12
felodipine with PVP, PVP/VA, HPMCAS, PVAcNH at 3294, 3298, 3348, 3354 cm⁻¹ against 3341 cm⁻¹50 wt% polymer filmskestur2010a R9
nimesulideNH at 3345 cm⁻¹ at every concentrationdichloromethane, 1–100 mg/mlkestur2011 R11
nimesulide with PVPPVP C=O ≈1684 → ≈1680 cm⁻¹, shoulder ≈1657 cm⁻¹20–50 wt% PVPkestur2011 R14
felodipine–PVP, nifedipine–PVPsecond PC with NH and carbonyl loadings10–90% and 25–70% PVP[rumondor2009a R7, R9]
naringenin, quercetin with PVPpolyphenol OH band 3138 and 3135 cm⁻¹; PVP C=O shift 21 and 24 cm⁻¹spin-coated films, 30% polyphenolwegiel2013a R2, R4
dextran–PVPPVP C=O 1680 → 1661 cm⁻¹; hydrogen-bonded fraction 1.00 (DEX1–PVP12) to about 0.09 (DEX450–PVP90) at 0.8 dextranspin-coated films[vaneerdenbrugh2012a R2, R14]
benzamide, phenacetin crystalsamide–amide only 100 and 94% alone; 0% with a COOH partner; 14 and 33% with C(O)NC2CSD survey[vaneerdenbrugh2011a R14]
lumefantrine–PVPVAcarbonyl bands fixed at 1736 and 1684 cm⁻¹ at 10, 25 and 50% drugdried spin-coated filmshiew2022a R17

Evidence table: hydrogen bonding.

Pitfalls and disagreements

Assignments rest on analogy with liquid models: the same non-bonded acid C=O is assigned at 1735 cm⁻¹ (chain ends) and 1726 cm⁻¹ (PVP-bonded molecule), and how one turns into the other is not shown taylor1997 C32, X4. IR bonds were recorded at 20–50 wt% polymer, whereas the growth effects were measured at 0.5–5 wt% and 70–110 °C kestur2010a X5; kestur2011 X10. The felodipine–PVP carbonyl shift is 25 cm⁻¹ in one paper and about 20 cm⁻¹ in a cited study marsac2009 R12.b; kestur2011 R20. A drug without a donor (ketoconazole) shows no shift yet mixes about athermally marsac2009 R7.c, R12.h. For bifonazole the donor–acceptor picture rests on crystal-structure reasoning and was not measured kestur2011 X9. The bond between indomethacin and PVP is proposed as the reason PVP inhibits crystallization, but no crystallization was measured in that paper taylor1997 X8. The PVP shoulder at 1661 cm⁻¹ with naringenin overlaps the drug's own band, and HPMC and PAA are placed on the IR trend line by projection wegiel2013a X9, X12. The bonded fraction of dextran–PVP rests on references set by one system and on films prepared differently from the DSC samples [vaneerdenbrugh2012a X12, X8]. No hydrogen bond was measured in the 15-drug screen, and its melt spectra are not shown [vaneerdenbrugh2011a X13].

← Concepts and formulas

concept · as used in Crystallization tendency

integration-controlled crystal growth

Crystal growth limited by the incorporation of molecules into the lattice at the surface, not by their transport to it; in the empirical law R_G = k_G S^{g} an overall order g of 1 means diffusion control and 2 integration control, with measured orders usually between 1 and 2 [alonzo2012 C3, E2]. Felodipine grew with g about 1.5 (a mix of the two) and with any HPMC level at g about 2, read as a shift toward integration control ("This suggests") [alonzo2012 R13, C32, C33].

Findings

F10.

Related concepts

contrasts with diffusion-controlled growth (named in [alonzo2012 C3]); related to crystal growth rate and step pinning.

Formulas

power-law crystal growth rate.

How it is measured or obtained

The slope of \ln R_G against \ln S from seeded growth rates over S 3–12, with \ln k_G from the intercept (or from a fixed order) [alonzo2012 P8, E3].

Values in the library
materialvalue or classconditionspaper ids
felodipine alone; with 0.2–3.5 µg/mL HPMCg about 1.50 ± 0.04; 1.85–2.17, mean 1.97 ± 0.12S 3–12, 25 °C[alonzo2012 R13]
felodipine, \ln k_Gabout −4.24 without; −5.42 to −5.86 with HPMC (fits with g fixed at 1.5 and 2)same[alonzo2012 R14]
absorptive dissolution test with residual crystals (Bioperformance)growth order about 1.7, agrees with the n of about 2 for integration control with an adsorbed polymerseeded[hate2019 C44]
Pitfalls and disagreements

The reading rests on the empirical meaning of g; step pinning and surface kinetics were not observed, the growth rates behind three of the six HPMC levels are not shown, and the k_G values come from fits with different fixed orders [alonzo2012 C34, X10, X8].

← Concepts and formulas

concept · as used in Crystallization tendency

interfacial free energy

The free energy per unit area (or length of a step edge) of the interface between a crystal or nucleus and the solution; a lower value gives a smaller critical nucleus and faster nucleation or growth, and the authors use it to explain why SDS speeds felodipine growth [schram2016 E3, C36]. New aspect (Physical stability): more generally, the free energy cost per area of creating an interface between a new phase (a nucleus or crystal) and its surroundings; in classical nucleation theory the thermodynamic barrier to nucleation grows with its cube konno2006 E8, C33; baird2012 E2. Additives can raise it (a polymer layer at a nucleus) or lower it (surfactants) [konno2006 C39; schram2016 C36]. Crystal–solution interface (formerly the page "crystal-solution interfacial free energy", from Crystallization from solution): the free energy per unit area of the boundary between a crystal or a crystal nucleus and the surrounding solution, written γ; in classical nucleation theory it enters the nucleation rate as \gamma^{3} in the exponent, so a small change in γ changes the rate steeply [duong2022b E6; indulkar2018 E6]. In the solution-crystallization papers it is the quantity through which surface-active additives are proposed to speed nucleation: lecithin adsorbing on the growing nucleus ("possibly"), SDS hemimicelles on the crystal surface ("is hypothesized to play a role") and phospholipid digests ("it is postulated") [indulkar2018 C40; indulkar2019b C33; enright2020 C26]. This content comes from the papers of 2018–2022 in Dissolution / D. Crystallization from solution; the free energy per area of a three-dimensional nucleus is distinct from the edge free energy per length of a two-dimensional nucleus used for step pinning (the felodipine edge value below). Interfacial tension (formerly the page "interfacial tension", from Crystallization from solution and Supersaturation from ASD dissolution), the name several papers use for the same quantity: the free energy per area of the interface between a crystal (or a crystal nucleus) and the solution, γ. It enters classical nucleation theory through the barrier \Delta G^{*} and the Kubota–Mullin effectiveness factor α through the surface excess energy [alonzo2012 E7; ilevbare2013c E4]. A surfactant that adsorbs at the crystal–solution interface lowers it, which can speed growth when this outweighs blocking of kink sites [ilevbare2012a C51]. The polymer is said to lower the surface tension needed to form the drug-rich phase of danazol, and SDS to lower the interfacial energy of felodipine; neither was measured [jackson2016 C36; schram2016 C36]. The danazol statement concerns the boundary of a liquid drug-rich phase, not of a crystal (synthesis: the drug-rich phase forms by liquid–liquid phase separation). New aspect (Supersaturation from ASD dissolution): the interfacial tension between an embryonic crystal and a water-saturated supercooled drug liquid is expected to be lower than against dilute aqueous solution, which together with the higher drug concentration favours nucleation in drug-rich droplets ("not unreasonable") [ilevbare2013d C40]. It enters the printed nucleation rate of the felodipine study as γ [konno2008a E3].

Findings

Crystallization from solution: F18, F22, F24, F34, F38, F39, F42. Physical stability: F4, F17. Supersaturation from ASD dissolution: F4, F13.

Related concepts

related to critical radius of a two-dimensional nucleus; general form of crystal-melt interfacial free energy; part of classical nucleation theory; sets the critical nucleus; related to kinetic barrier to nucleation, hemimicelle, mixed micelles, polymer-surfactant interaction, heterogeneous nucleation and drug-rich phase.

Formulas

critical radius of a two-dimensional nucleus (formula table only); classical nucleation theory, classical nucleation barrier, pre-exponential factor of classical nucleation theory, Kubota-Mullin impurity effectiveness factor. The critical nucleus radius r^{*} = 2\gamma\nu/(kT\ln S) follows from γ duong2022b E9.

How it is measured or obtained

For felodipine the edge free energy γ = 3.9 × 10⁻²⁰ J/nm came from water and ethylene glycol contact angles (Wu harmonic mean, angles not printed) [schram2016 P9, R13, X10]. Not measured in the solid-state physical-stability papers, where it enters arguments only konno2006 C39. In solution, values follow from linearized nucleation-rate data duong2022b E7. The crystal–solution value is not measured directly in the library. One paper obtained it from the slope of \ln(J/S) against 1/\ln^{2}S fitted to induction times converted into nucleation rates, with a molecular volume of 920 ų and T = 310 K, for posaconazole in buffer and three intestinal media duong2022b P15, E7. The other papers infer a change of γ from shorter induction times, from pyrene signals of adsorbed surfactant, or from a cited surface tension of a medium (32.6 against 54.4 mN/m) [indulkar2019b R6, R7; enright2020 R32]. Not measured in the crystallization-from-solution papers that call it interfacial tension. For paclitaxel the crystal–water value could not be determined and literature values of representative compounds were used [que2018 C37]. Surface tension measurements were used only to find polymer adsorption on seeds [alonzo2012 P6]. Not measured in the ASD dissolution papers either [ilevbare2013d C40].

Values in the library
materialvalue or classconditionspaper ids
felodipine crystal edgeγ 3.9 × 10⁻²⁰ J/nm (contact angles, not printed); ρc 3.98 nm at S 425 °C[schram2016 R13, R14, X10]
atazanavir in lecithin media; with lecithin, lyso-PC and oleatelecithin "possibly" lowers the interfacial energy between solute and emerging crystal; proposed to be lowered, more by the digests; not measured3 mM STC, pH 6.5; bile salt media, pH 6.5[indulkar2018 C40, E6; enright2020 C26]
felodipine with polymera polymer-rich layer at the nucleus "may also increase" the surface penaltyargumentkonno2006 C39
felodipine solution with SDSfaster growth explained by a lower interfacial energyS = 4[schram2016 C36]
felodipine–PVPVA–TPGSTPGS lowering the energy between separated phases slows coarseningASD hydration[saboo2021 C40]
flutamide prenucleiunfavourable interfacial term outweighs the volume term below a critical radiusframeworktrasi2012b C46
posaconazole, PBS pH 6.53.52 mJ/m²37 °C, fit of the high-supersaturation induction timesduong2022b R16
posaconazole, FaSSIF V1, FaSSIF V2, FeSSIF V12.61, 2.73, 3.24 mJ/m²; FeSSIF V2 not fittableas aboveduong2022b R16, R15
naproxen, acetaminophen, tolazamide, lovastatin, cefodizime sodium, prasugrel hydrochloride, spironolactone (cited)0.60–2.80 mJ/m²literatureduong2022b R18
atazanavir with SDS hemimicellesproposed to be lowered; not measured0.075–0.3 mg/mL SDS[indulkar2019b C33, X4]
felodipine–HPMCγ, a and L of eq 7 not givenKubota–Mullin[alonzo2012 C27]
surfactants on ritonavirgrowth about twice as fast with TPGS or Tween 80 at 5 µg/mLseeded, S 7.6[ilevbare2012a R17, C51]
paclitaxel critical radius 5–7 nmfrom literature surrogates of γS 88[que2018 R15, C37]
supersaturated drugs in surfactant mediasurfactants should lower γ and speed nucleation; human fluid slowed it insteadclassical nucleation theory, 37 °C[elkhabaz2021 C20, C21]
Pitfalls and disagreements

The interfacial energy in SDS or HPMCAS solution was not measured [schram2016 X8]. The solid-state papers use homogeneous theory although nucleation in the films is "most likely heterogeneous" konno2006 C33. The papers disagree on whether γ carries the medium effect: the posaconazole fits find no trend of induction time with γ and assign the effect to the pre-exponential factor, whereas the lecithin, SDS and digest papers assume a lower γ [duong2022b C51, C52; indulkar2018 C40; indulkar2019b C33]. The fits rest on 3–9 points per medium, and γ also falls in the induction-time order for three of four media, so a γ contribution is not excluded duong2022b X11, X12. The human-fluid paper notes that surfactants that lower γ should speed nucleation, while human fluid slowed it, and adds an attachment-rate term [elkhabaz2021 C20, C22]. In the celecoxib work on polymer combinations the differences between polymers were not attributed to interfacial tension, because polymer adsorption on the nucleus would lower it and raise the nucleation rate, the opposite of what was observed; the point is argued, not measured xie2015 C43. This is opposite in sign to the felodipine argument that a polymer-rich layer at the nucleus "may also increase" the surface penalty konno2006 C39 (synthesis: two papers assume opposite effects of polymer on γ; neither measured it). The lower interfacial tension in droplets is an expectation, not a measurement [ilevbare2013d C40]. Interfacial tension with and without polymer was not measured, and the text cites the wrong equation for it xie2015 C43, X1. The same γ is reported per area (mJ/m²) for three-dimensional nuclei and per length (J/nm) for the step edge of a two-dimensional nucleus, so the two kinds of value are not comparable [schram2016 R13; duong2022b R16] (synthesis: different units and geometry).

← Concepts and formulas

concept · as used in Crystallization tendency

intramolecular hydrogen bonding

A hydrogen bond between two groups of the same molecule

It matters for polymer selection because it removes a donor or acceptor from the pool available to the polymer: the sulfonamide NH of nimesulide bonds to its own ether oxygen, which the authors say "considerably diminished" its capacity to bond to polymers and "most likely" explains why the polymers differ less than anticipated kestur2011 C20, C27, C28. Flufenamic acid holds its aniline N–H in a resonance-assisted hydrogen bond (N···O 2.682 Å in form III and 2.646 Å in form I, against a range of 2.66–2.74 Å for such bonds) and is therefore regarded as interacting only through its acid group vaneerdenbrugh2010b R12, C12. Further cases: the 5-OH and keto group of naringenin and the 5-OH and 3-OH of quercetin, which leave two and three phenolic OH groups free for the polymer wegiel2013a M1, M2, C30; the aniline N–H of flufenamic acid (DFT) and the energy profile of anthranilic acid, where one aniline hydrogen stays free [vaneerdenbrugh2011a R5, R6, C18]; and the OH of lumefantrine, bonded to its own nitrogen in the crystal hiew2022a R33, C61.

Findings

Miscibility and drug-polymer interactions: F14, F15, F17, F24, F25, F26. Physical stability: F18, F24.

Related concepts

part of hydrogen bonding; limits drug-polymer interactions and hydrogen-bond donor and acceptor groups; contrasts with self-association; makes acid-base interaction the alternative wegiel2014 C44.

Formulas

none.

How it is measured or obtained

By a donor band that stays at the same wavenumber on a 100-fold dilution in a non-polar solvent (nimesulide NH at 3345 cm⁻¹, 1–100 mg/ml); by no change on adding a strong acceptor or donor (1-ethyl-2-pyrrolidinone, acetic acid); by comparing the band of the amorphous and crystalline forms; by crystal structure and DFT geometry kestur2011 R10, R11, R12, R17, R19. For flufenamic acid, from N···O distances in two crystal structures vaneerdenbrugh2010b P11, R12. By a conformer energy profile of the single molecule in vacuum trasi2012b P9, R24; by an IR band of the amorphous drug that matches the intramolecularly bonded form of the crystal wegiel2014 R2, C11.

Values in the library
materialvalue or classconditionspaper ids
nimesulideNH 3345 cm⁻¹ at 1–100 mg/ml in dichloromethane; unchanged with added acid or lactamIR, 100 µm cellkestur2011 R11, R17
nimesulideNH ≈3288 (crystal), ≈3276 cm⁻¹ (amorphous)KBr; spin-coated filmkestur2011 R12
flufenamic acidN···O 2.682 Å (form III), 2.646 Å (form I)crystal structuresvaneerdenbrugh2010b R12
flutamide; curcumin (other chapters)intramolecular CH···O leaves a poor acceptor; intramolecular bond limits polymer bondingcalculation; IR, stabilitytrasi2012b C35; wegiel2014 C3
naringenin, quercetin5-OH to keto O (both); 3-OH to keto O (quercetin); PVP still bonds the free OH (OH band 3138 and 3135 cm⁻¹)structure; IRwegiel2013a M1, M2, R2
anthranilic acidenergy about 31–33 kJ/mol up to 30° (acid O–H to N), about 50 at 80–90°, about 0 at 180° (N–H to carbonyl O); no bifurcated bondDFT dihedral scan[vaneerdenbrugh2011a R6, C18]
lumefantrineOH···N bond in the crystal; no PVPVA carbonyl shiftcrystal structures (cited); IRhiew2022a R33, R17
flutamideplanar lowest-energy conformer with the carbonyl next to C(5)–H; second minimum ≈2.7 kJ/mol higher; barriers ≈23 kJ/molDFT, vacuumtrasi2012b R23, R24
flutamide crystalamide carbonyl in a bifurcated bond, one contact intramolecularcited structuretrasi2012b R26
amorphous curcuminketo–enol band at 1626 cm⁻¹ as in the crystal; no carbonyl band near 1700 cm⁻¹IRwegiel2014 R2, C11
curcumin against resveratrol with PVPbonded PVP carbonyl dominant only at 60% curcumin, already at 50% resveratrolIR of filmswegiel2014 R8, R9, C32

Evidence table: intramolecular hydrogen bonding.

Pitfalls and disagreements

For the amorphous state the evidence is that a broad band lies near the crystal's, and the crystal contains an intermolecular NH bond too kestur2011 R19, R12. The NH maximum shifts as much with PVAc as with PVP in the films, which weakens the argument that the intramolecular bond alone sets the polymer ranking kestur2011 X16. The treatment of flufenamic acid as purely acidic is an assumption supported by crystal geometry, not by a spectrum in the films vaneerdenbrugh2010b C12. An intramolecular bond limits but does not exclude polymer bonding when other donors remain (naringenin and quercetin bonded PVP strongly) wegiel2013a R2, R4. Whether the lumefantrine OH bond persists in the dispersion was not measured hiew2022a C61. No curcumin analogue without the intramolecular bond was tested, and within the curcumin paper PVP still gave the largest shifts wegiel2014 X13. For flutamide the bond in the glass is inferred from a vacuum calculation, and PAA's failure may reflect poor mixing instead trasi2012b C35, X12.

← Concepts and formulas

concept · as used in Crystallization tendency

kinetic barrier to nucleation

The activation energy for moving molecules across the nucleus–matrix interface, \Delta G_a in classical nucleation theory, related to diffusivity or reciprocal viscosity; distinct from the thermodynamic barrier set by interfacial energy and driving force konno2006 E8, C33; marsac2006a E9, C31. Experimentally the Arrhenius activation energy of the nucleation rate is used as its measure marsac2006a R3, E2.

Findings

F1, F2, F4.

Related concepts

part of classical nucleation theory; contrasts with thermodynamic driving force for crystallization; related to molecular mobility and polymer rejection at the crystal growth front.

Formulas

classical nucleation theory, Arrhenius equation.

How it is measured or obtained

Not measured directly; nucleation rates at several temperatures give an Arrhenius slope, taken separately above and below Tg marsac2006a P5, R2, R3; otherwise argued from the size of a polymer effect at low content konno2006 C40, C41.

Values in the library
materialvalue or classconditionspaper ids
nifedipine, felodipineactivation energy below Tg 122 (11) and 145 (3) kJ/mol; above Tg 78 and 95Arrhenius, ≈25–75 °Cmarsac2006a R3
with 3 wt% PVPbelow Tg 127 and 201; above Tg 103 and 111 kJ/molsamemarsac2006a R3.c, R3.d
felodipine with 3–25% PVP, HPMCAS, HPMCspeculated rise of ΔGa, set by amount rather than type of polymer22 °C, drykonno2006 C40, C41

Evidence table: kinetic barrier to nucleation.

Pitfalls and disagreements

The Arrhenius plot carries no rate unit and its temperatures are given only by the axis; the activation-energy difference between the two drugs is significant only below Tg, a restriction the abstract drops marsac2006a X2, X6. The polymer-raised barrier is speculation ("it seems reasonable to speculate") konno2006 C40.

← Concepts and formulas

concept · as used in Crystallization tendency

kinetic stabilization

Stabilization of an amorphous drug against crystallization by slowing the process (molecular mobility, nucleation, interfacial factors) while the thermodynamic driving force stays large. Marsac and co-workers conclude from a small predicted reduction of the driving force, a low predicted solubility in PVP and the faster nucleation of the pure drugs that inhibition "can therefore be attributed predominantly to a kinetic stabilization"; molecular-level miscibility is "of critical importance" because it alters "thermodynamics and, more importantly, dynamics" marsac2006b C41, C42, C55, C56, C59. The same group states that thermodynamically stable dispersions are hard to achieve and that apparent stability is "most likely achieved through modification of the kinetics" marsac2009 C34. For the growth of felodipine with PVP the effect "is believed to be kinetic rather than thermodynamic" kestur2010b C28.

Findings

F9.

Related concepts

contrasts with thermodynamic driving force for crystallization; depends on drug solubility in polymer; related to crystallization inhibition and amorphous solid dispersion.

Formulas

Hoffman equation.

How it is measured or obtained

Not measured directly in this chapter. It is inferred by elimination: the thermodynamic estimates (Hoffman driving force, predicted solubility) are small or low, while the dispersions stay amorphous marsac2006b R12, R17, X12; marsac2009 R11, X17. Kinetic quantities (nucleation rates, mobility) are cited from a companion study or not measured marsac2006b R16, C59.

Values in the library
materialvalue or classconditionspaper ids
nifedipine with 40% PVPdriving force −7.95 against −9.0 kJ/mol pure at 20 °CHoffmanmarsac2006b R12
nifedipine, felodipine in PVPpredicted solubility 0.06–0.07 and 0.25–0.31 g/gFlory–Hugginsmarsac2006b R17
five compounds in PVP K29/3282–99% of the drug above solubility at 0.50 polymerpredictedmarsac2009 R11
pure nifedipine and felodipine against their dispersionsnucleation much faster for the pure drugscitedmarsac2006b R16

Evidence table: kinetic stabilization.

Pitfalls and disagreements

The kinetic reading is an inference: no crystallization, nucleation or mobility data are reported in either paper marsac2006b X12; marsac2009 X17. Being below the estimated solubility did not make PEG dispersions stable, which points to nucleation by the carrier as another route zhu2012 X13. A small dissolved fraction may explain why crystallization of a dispersion is not always complete marsac2009 C35.

← Concepts and formulas

concept · as used in Crystallization tendency

kinetic versus thermodynamic control of crystal growth

Growth of a crystal from a supercooled liquid needs both mobility and a driving force: near Tm mobility is high and the driving force low (thermodynamic control), and on cooling the driving force rises while mobility falls (kinetic control) mosqueragiraldo2014 C25. Felodipine growth that slowed on cooling over 70–110 °C was read as transport-limited kestur2010b C22, C23. In the indomethacin polymorph paper mobility limits growth below the temperature of fastest growth and the driving force above it, and the steep rise of polymer inhibition above that temperature is placed in the second region tian2017 C22, C35, C36.

Findings

F3, F22, F29.

Related concepts

depends on thermodynamic driving force for crystallization and molecular mobility; related to melting point depression and degree of supercooling; contrasts with Tg-scaled crystal growth rate.

Formulas

temperature of maximum crystal growth rate.

How it is measured or obtained

From where growth peaks between Tg and Tm and from how an additive's effect changes with temperature trasi2012a C35; mosqueragiraldo2014 C36.

Values in the library
materialvalue or classconditionspaper ids
celecoxib with 10% TPGS and 10% PVPcelecoxib Tm 151.49 °C (12 °C depressed); growth about 0.03 of PVP alone at 120 °C, 0.6 at 70 °Cseeded melt growthmosqueragiraldo2014 R5.f, R2.b, X5
celecoxib with 3, 5, 7% PVP, with and without TPGSpaired curves cross at 110, 100, 90 °Csamemosqueragiraldo2014 R3, R4, C14
felodipine form IIgrowth falls on cooling from 105 to 70 °C although the driving force risesmelt filmskestur2010b C22, C23
acetaminophengrowth maximum closer to Tm, nucleation at larger supercoolingclassical picturetrasi2012a C35
indomethacin α with 10% HPMCfastest growth moved from 135 to 100 °C; ratio without/with polymer ≈11 (100 °C) → ≈2900 (120 °C)melt growthtian2017 R15, R12
Pitfalls and disagreements

The driving force was not calculated for the celecoxib mixtures, and melting-point depression alone does not order them mosqueragiraldo2014 C33, X13. The celecoxib growth temperatures were assumed to lie in the mobility-controlled range mosqueragiraldo2014 C26.

← Concepts and formulas

concept · as used in Crystallization tendency

liquid-liquid phase separation

The splitting of a highly supersaturated aqueous drug solution into a water-rich continuous phase and a dispersed, non-crystalline drug-rich phase, before crystals form; the authors call it glass–liquid phase separation when the drug-rich phase is a glass ilevbare2013a C2; indulkar2015 C38. The concentration at which it starts is called the LLPS concentration, CL-L or coexistence concentration, and in this chapter equals the amorphous solubility within the accuracy of the methods ilevbare2013a C14, C3. It was earlier discussed as "promiscuous aggregation" in screening assays and as "oiling out" in crystallization ilevbare2013a C7, C9, C35. The papers of 2017–2022 add a weak base shifted from simulated gastric fluid to pH 6.5: pazopanib gave a milky, bluish solution at once, with particles of 495.9 nm (145.4 nm with 50 µg/mL HPMC) and no particles at a dose below its amorphous solubility [sugihara2018 R14, R15, R20], and ibuprofen droplets that measured 3182–5313 nm in buffer and 268–345 nm with HPMC or PVP-VA [ueda2020b R5]. New aspect (Bioperformance): the two phases are in metastable equilibrium with equal activity [raina2014a C3, C4, C33; indulkar2016 C1; hate2018 C5]. For high-Tg drugs such as telaprevir the term glass–liquid phase separation is used [mosqueragiraldo2018b M1]. Drug-rich spheres were also found by SEM in pooled human intestinal fluid, the first such demonstration according to the authors [elkhabaz2019b R4, C24]; a crystalline co-drug lowers the concentration at which atazanavir separates [alkalla2026 C2]. New aspect (Supersaturation from ASD dissolution): dispersion dissolution, unlike dissolution of a neat amorphous drug, can push the concentration past the amorphous solubility, so that LLPS follows dissolution and precedes crystallization [jackson2015 C7, C29]; the colloids of rapidly dissolving ritonavir dispersions "highly likely" originate from LLPS [ilevbare2013d C33].

Findings

Amorphous solubility and LLPS: F1, F2, F3, F4, F5, F8, F9, F11, F13, F15, F16, F22, F23. Bioperformance: F1, F2, F4, F5, F10, F18, F19, F27. Supersaturation from ASD dissolution: F7, F9, F10.

Related concepts

produces the drug-rich phase; special case glass-liquid phase separation; sets the amorphous solubility; described by the binodal and spinodal decomposition; grows by droplet coalescence; precedes crystallization (a precursor, F3). Produces drug-rich nanodroplets; related to metastable equilibrium.

Formulas

binodal condition, amorphous solubility advantage with water-sorption correction, ideal-mixing reduction of amorphous solubility.

How it is measured or obtained

By extinction at a non-absorbing wavelength during slow addition of a drug stock, by a polarity-sensitive probe or the drug's own fluorescence, by DLS or NTA sizing of the droplets, by ultracentrifugation of the phases, by NMR of dissolved drug, by a flux break, and by a solvent-free pH-shift titration for weak bases [ilevbare2013a P5, P10, P11; almeidaesousa2015 P1, P2; mosqueragiraldo2015 P10; trasi2015a P6; indulkar2015 P10] (F4). In the Bioperformance papers: rise of extinction on solvent shift; turbidity without birefringence under polarized light raina2014a R4, C12; droplet size and number by NTA or DLS [indulkar2016 P4; hate2018 R6, R7]; the 369 nm fluorescence band of atazanavir [hate2018 P6, C18]; the flux plateau; antisolvent titration read by 450 nm extinction [alkalla2026 P7, C16]; the drug's own fluorescence intensity and SEM of the centrifuged pellet [elkhabaz2019b R3, R4, C17]. During dispersion dissolution: the pyrene ratio, with the onset read as the UV concentration at the pyrene drop; stepwise dissolution with dilution; turbidity titration with polymer present [jackson2015 P3; ilevbare2013d P4, P2].

Values in the library
materialvalue or classconditionspaper ids
ritonavir dropletsabout 345 nm at 1 min, about 842 nm at 64 minwater, 37 °C, single addition at the onsetilevbare2013a R3
ritonavir drug-rich phase94.85–95.97% drug, 4.03–5.15% water, Tg 0.4 °C10–50 °C, ultracentrifugedilevbare2013a R5, R6
weak-base dropletsZ-average about 116–387 nmclotrimazole, atazanavir, nicardipine, 37 °Cindulkar2015 R14
telaprevirspecies about 300 nmpH 6.8, 37 °Cmosqueragiraldo2015 R5
ritonavir dropletsabout 360 nm at 30 µg/mL100 mM pH 6.8, 25 °C[trasi2015a R15]
pazopanib colloids495.9 nm without, 145.4 nm with 50 µg/mL HPMC (DLS); NTA about 50–650 nmpH 6.5 after pH shift, 37 °C[sugihara2018 R15, R16]
ibuprofen droplets3182–5313 nm (buffer); 272–345 (HPMC); 268–320 (PVP-VA); 472–808 nm (EUD-E)pH 2.0 (D2O), 37 °C, fresh[ueda2020b R5]
felodipineonset 5.70–13.4 µg/mL20–45 °C, bufferraina2014a R1
nifedipineLLPS only with 1 mg/mL HPMC: 30.86–88.89 µg/mL20–45 °Craina2014a R1, R5
clotrimazoleonset 7.5 µg/mL; droplets 111–257 nmpH 8[indulkar2016 R2, R4]
atazanavir78 µg/mL; droplets 204–226 nmpH 6.8[hate2018 R2, R6]
ritonavironset about 38 µg/mL in waterUV extinctionilevbare2013a R2
enzalutamideonset about 42 µg/mLpH 6.5, 37 °C[wilson2018 R2, C16]
atazanavir, darunavir93.1 and 363.8 µg/mL (antisolvent, 450 nm)pH 6.8, 37 °C[alkalla2026 R3, C16]
pretomanidabout 166 µg/mL in buffer, 308 in FaSSIFpH 6.5, 37 °Cnguyen2023a P2, R2
danazol dispersions (eight)onset about 6.0–10.4 µg/mL against 8 µg/mLpH 6.8, 25 °C[jackson2015 R5, M1]
ritonavir–PVP dispersion, stepwiseonset between 10 and 20 µg/mL37 °C[ilevbare2013d R1, X8]
ritonavir, 180 µg/mL PVP / PAA / HPMC / HPMCAS / CAAdP 0.8518.3 / 18.3 / 18.7 / 18.2 / 21.3 µg/mL (18.2 without)titration, 37 °C[ilevbare2013d R10]
Pitfalls and disagreements

Extinction alone cannot tell droplets from crystals, and a fast crystallizer may crystallize before it phase separates (griseofulvin without polymer) almeidaesousa2015 C16, C18. The ritonavir phase diagram rests on four binodal points between about 11 and 29 °C ilevbare2013a X17. Droplet size grows with time by coalescence and varies with drug and technique ilevbare2013a R3, C18. In SDS the 369 nm band that marks the atazanavir drug-rich phase in buffer appears from about 200 µg/mL, far below the amorphous solubility of about 1450 µg/mL, so a spectral marker has to be checked in the micellar medium [indulkar2017a R17, R18, C27]; extinction rose on the first addition to STCDC and SGCDC without visible turbidity [lu2017b R4]. LLPS is seen only if nucleation is slow compared with phase separation; the metastable zone for LLPS is "small" by the authors' reading raina2014a C30, C32. Turbidity does not tell LLPS from crystallization without a crystal-specific method [elkhabaz2017 C30]. Above the onset an inhibitor can still fail: pretomanid with HPMCAS-HF crystallized within minutes in the presence of droplets (F27) nguyen2023a R7, C13. A polymer in the droplets lowers the onset (F19) [ueda2020a R1, C13]. In the danazol dispersion study the overshoot above the amorphous solubility before LLPS was inferred from UV, which is unreliable there [jackson2015 X12]; a slowly releasing dispersion (50:50 HPMCAS danazol) never reached it [jackson2015 C33].

← Concepts and formulas

concept · as used in Crystallization tendency

matrix crystallization

Crystallisation of the amorphous solid itself while it is in contact with the medium, before the drug has dissolved (also called solid-mediated crystallisation), in competition with dissolution and with crystallisation from solution [moseson2021 C1, C27]. Bicalutamide–PVPVA is the case in point of the ASD process papers: a hydrated, polymer-depleted, drug-rich solid that is "likely" below its Tg at 37 °C becomes mobile enough to crystallise in place, a surface-driven reaction triggered by hydration and polymer loss [moseson2021 C55, C56, C59]. Moseson (2022) tested the route across loadings: bicalutamide–PVPVA at 5 and 10% loading released drug and polymer together and escaped it, whereas 20–50% loading, cryomilling and cryomilled surfactant-containing powders raised the risk. The authors call it surface-mediated: a polymer-depleted, drug-rich boundary layer forms, crystallizes and "could prevent release" moseson2022b C36, C39, C43, R6, R7. In posaconazole–HPMCAS at 50% loading a 12 nm coating went with a much lower SHG signal after 3 h and higher release, which the authors suggest is a delay of matrix crystallization moseson2023 R15, R18, C40. New aspect (Bioperformance): crystallization inside the undissolved ASD particle during dissolution, as against crystallization from solution [mosqueragiraldo2018b C40; elkhabaz2017 C34]. The poor exposure of the 10% enzalutamide–HPMCAS dispersion was attributed to rapid matrix crystallization on contact with water; the printed evidence is crystals in the medium and a 10 µg/mL maximum [wilson2018 C56, X11]. New aspect (Supersaturation from ASD dissolution): in the first study of the two routes it is called "crystallization from matrix": the amorphous particle converts to crystal on contact with the medium, against nucleation and growth from the supersaturated solution alonzo2010 C43, C47. If it is faster than dissolution, no concentration above that of the crystal can be reached, because the crystal is what dissolves alonzo2010 C33. The papers of 2015–2018 show that the dispersion polymer and its share decide it: resveratrol dispersions needed 80–95% cellulosic polymer to stay amorphous in buffer, PAA failed for celecoxib "irrespective of the drug loading", and an inhibitor dissolved in the medium cannot reach it [wegiel2015 R14; xie2016a C33, C31; xie2015 C35].

Findings

ASD process: F27, F28, F29, F38, F39, F40, F41, F46. Bioperformance: F9, F11, F12, F23, F26, F27, F30. Supersaturation from ASD dissolution: F1, F2, F3, F16, F24, F27.

Related concepts

contrasts with desupersaturation; accelerated by residual crystallinity; not blocked by crystallization inhibition by polymers in solution; accompanies incongruent release and surface drug enrichment; related to surface crystallization; avoided by congruent release; fed by the drug-rich surface layer; delayed by an atomic layer coating in one posaconazole system. Contrasts with solution crystallization; favoured by drug loading; part of crystallization during dissolution; related to plasticization by water.

Formulas

None specific; the competition with dissolution is set out with the Noyes-Whitney equation alonzo2010 E3.

How it is measured or obtained

Time-lapse polarised light microscopy of particles in buffer, in-line Raman of a slurry resolved by MCR-ALS, XRPD and SEM of recovered solids, polymer release by size-exclusion chromatography, and dissolution at several doses read for plateaus below the amorphous solubility and a falling fraction dissolved [moseson2021 P14, P15, P16, R17, R18, R19]. SEM, SEM-EDX (fluorine occurs only in the drug, F/F0) and confocal fluorescence of compacts exposed face down to medium for 5–60 min; Raman slurries resolved by MCR-ALS for the form 2 time course moseson2022b P12, P14, P15, P16, P17. SHG of particles after 3 h in buffer moseson2023 R18. Polarized light microscopy of particles in buffer [mosqueragiraldo2018b R18]; SHG [elkhabaz2017 P10]; indirectly, a flux ceiling not reached [mosqueragiraldo2018b R16]. In the ASD dissolution papers: in situ Raman of aqueous slurries with a PLS model for crystallinity or marker bands, PXRD of wetted powder after 15 min, and polarized light microscopy of wetted particles alonzo2010 P5, P6, P8; raina2014b P4, P6, P10; the 1630/1605 cm−1 Raman ratio of resveratrol slurries, also in a medium where the enteric polymer does not dissolve (pH 1.2) wegiel2015 P7, A3; the amorphous Raman shoulder near 1598 cm−1 of celecoxib slurries xie2015 P3, R3; polarized light of residues retrieved after the release peak [xie2016a P6].

Values in the library
materialvalue or classconditionspaper ids
BCL–PVPVA 30:70, crystal-freeform 2 begins at 20–30 min (half conversion about 36 min); plateau about 37–38 µg/mL against an amorphous solubility of 150 µg/mL60 and 150 µg/mL doses, pH 6.8[moseson2021 R12, R13, R19]
BCL–PVPVA with residual or added form 1 crystalsform 1 first detectable at about 15 min (S/155), 7 min (M/155), from the first spectrum (10% seeds)Raman slurry, 10 mg/mL[moseson2021 R21]
amorphous nifedipine (outside)crystallises mainly by the matrix route within about 10 min; PVP, HPMC or HPMCAS in solution do not change onset or routebuffer, 25 °Craina2014b C10, C11
celecoxib dispersions (outside)polymers dissolved in solution cannot inhibit crystal formation in the matrixhigh drug loadingxie2015 C35
telaprevir 50% HPMCAS (outside)slow, incongruent release; HPMCAS leaves about twice as fast as the drug, leaving a drug-rich layeroptical micrographs[mosqueragiraldo2018b C40, C43]
enzalutamide 10:90 HPMCAS (outside)rapid matrix crystallisation on contact with waterdissolution[wilson2018 C56]
posaconazole ASDs in acid (outside)matrix crystallisation about 68% at 50% DL, about 0.05% at 25% DLSHG over TPE-UVF, 30 min[elkhabaz2019a R20, C38]
BCL–PVPVA 5 / 10 / 20 / 30 / 50% DL powdersend level about 91 / 96 / 88 / 86 / 73% of the dose; polymer ahead of drug from 20% DLdose of at most 30 µg/mL, pH 6.8moseson2022b R6, R7
30% and 50% DL compacts, 1 hdrug-rich boundary layer of 94 ± 8 µm with crystals (F/F0 1.41 ± 0.10); 5–10 µm crust of crystalsSEM-EDXmoseson2022b R15
30% DL cryomilled 2 and 5 minabout 50 and 49% of the dose, plateau about 15 µg/mLsamemoseson2022b R4
10% DL after 10 min of cryomilling or extrusion and 30 s of millingabout 93% of the dosesamemoseson2022b R8
30% DL with 3% TPGS, cryomilled 5 minstops at about 60% (about 18 µg/mL)samemoseson2022b R12
50% PCZ–HPMCAS, small particles, uncoated against 12 nm Al2O3SHG 126.7 against 3.7; about 3.35 against 5.1 µg/mL at 3 hsingle stage, pH 6.5moseson2023 R18, R15
telaprevir 50% HPMCAS ASDbirefringent domains in the particle at 20 min; 0.65 µg/minpH 6.8[mosqueragiraldo2018b R16, R18]
telaprevir 47.5:47.5:5 SDS ASDnone visible; 1.07 µg/minsame[mosqueragiraldo2018b R16, R18]
ezetimibe PAA ASDcrystals within minutes, inferred from the matrixbuffer, FeSSIF[elkhabaz2017 C34, R12, R16]
posaconazole ASDs 10 / 25 / 50%SHG crystallinity 0.3–0.4 / 3–4 / 19–26%60 min FaSSGF[shah2024 R21]
posaconazole ASDs50% DL final crystallinity about 250 times that of 25% DL[elkhabaz2019a R18]
enzalutamide 10:90 HPMCAS-MF ASDpeak about 10 µg/mL, crystals in the mediumpH 6.5, 100 µg/mL total[wilson2018 R6, C56]
pretomanid HPMCAS-HF ASD tabletscrystalline drug in the particles after FeSSIF at pH 5.5 and below, and after a 1 h gastric stagePXRD and SEM, 30 mg in 150 mLnguyen2023a R20, R22
neat amorphous felodipine≈54% crystalline at 1 min (37 °C); 50% after about 8 min (25 °C)pH 6.8 slurryalonzo2010 R2
felodipine with 500 µg/mL PVP, HPMC, HPMC-AS in the medium50% crystalline at ≈20.5, ≈18.5, ≈13.5 min37 °Calonzo2010 R6
neat amorphous indomethacinα form present at 90 min, like pure α at 120 minpH 2, 37 °Calonzo2010 R12
nifedipine 50:50 dispersions (PVP shown)particles not birefringent; crystals form in the solutionpH 6.8raina2014b R13, X18
nifedipine–HPMCAS 50:50 at pH 2matrix amorphous for 3 hpolymer barely solubleraina2014b R15
celecoxib–HPMCAS 33.3% particles in pH 3 suspensioncrystals on or in particles after about 7 days (Triton X100), none in 48 h25 °C[chen2015a R20, R14]
resveratrol 35% with HPMC / HPMCAS / CMCABRaman ratio ≈0.87 / ≈0.84 / ≈0.81 (crystal 0.76) within hourspH 6.8, 37 °Cwegiel2015 R9
resveratrol, minimum polymer to prevent it80% HPMC, 85% HPMCAS, 95% CMCABpH 6.8, 37 °Cwegiel2015 R14
neat amorphous celecoxibcomplete within an hour; crystals on the particles from 2 minpH 6.8, 37 °Cxie2015 R4, R8
celecoxib–PVP-K12 50%crystalline within 4 h in a slurrypH 6.8, 37 °Cxie2015 R4
celecoxib–PAA 5:5 and 1:9, HPMCAS in the mediumrelease stopped at about half; needles in the residuepH 6.8, 37 °C[xie2016a R9, R10]
lopinavir 50% with EUD, CAP / HPMCPcrystals by 8–16 h / none at 48 hpH 3, 25 °C[li2018 R7]
delamanid salt dispersions, films after 1 h in acidclear at pH 1.6 (HPMCP); crystals at pH 3.0 on all but HPMCP-55; small crystals on every film at pH 5.025% drug, 37 °C[nguyen2023c R19]
same, powderspitted, porous HPMCAS surfaces with agglomerates increasing with pH; HPMCP-50 covered at pH 5.01 h in acid[nguyen2023c R18]
delamanid dispersions in HPMCP, films with and without a 2 µm Eudragit L 100-55 layerstar- and needle-shaped crystals on every uncoated film; none under the layerpH 3.0, 1 hnguyen2023b R3, R6
posaconazole–HPMCAS dispersionscrystals by SHG in every system after 30 min in FaSSGF, more numerous after transfer37 °C[elkhabaz2019a R16]
Pitfalls and disagreements

The hydrated-solid mobility mechanism was not measured (the wet Tg of amorphous bicalutamide could not be determined) and the kinetics come from slurries far above the dose concentrations [moseson2021 R27, X8, X11]. Attribution to the matrix route is often inferred: crystals could instead nucleate in the concentrated boundary layer around the dissolving particles, or the evidence is crystals in the medium and a low maximum without a solid-state analysis [xie2016a X6; wilson2018 X11]. Solution-mediated growth is not absent at high dose [moseson2021 X4]. The results and the discussion of the 2022 study disagree on whether 20–30% loading crystallized, the crystals of the compacts were identified by shape and not by diffraction, and the polymer-depleted state of the crust is inferred from a dye moseson2022b X14, X16, X18. The Tg and composition of the hydrated layer were not measured moseson2022b C60. One micrograph without XRPD supports the telaprevir case [mosqueragiraldo2018b X13]. Matrix and solution crystallization were not separated for enzalutamide: no solid-state analysis of the wetted dispersion is shown (F9) [wilson2018 X11]. In the ASD dissolution papers, slurries held far more solid per unit of polymer than dissolution vessels (about 500–750-fold for felodipine), so their lag times rank systems but do not give the time scale in dissolution alonzo2010 X5. The proposal that crystallization starts at the particle surface and that dissolved polymer delays it there was not tested alonzo2010 C36. For felodipine at 25 °C the fall after the peak may come from the matrix as well as from solution alonzo2010 X6. The PVP-K12 celecoxib dispersion crystallized in a slurry yet released fully in dissolution xie2015 R4, R6. PXRD found no crystals in delamanid dispersion powders after 1 h in acid, a method that detects about 1–5% crystallinity, so the crystal mass that harmed release was small [nguyen2023c R15, R16, C46].

← Concepts and formulas

concept · as used in Crystallization tendency

maximum crystal growth velocity

The highest crystal growth rate of a compound over temperature (MCV), reached at Tcryst,max between Tg and Tm; below Tcryst,max growth is limited by mobility, above it by the shrinking driving force trasi2014a C22. The MCV is said to occur near 0.94 Tm trasi2014a C37, E8.

Findings

F1, F2.

Related concepts

part of crystal growth rate; related to degree of supercooling and melt viscosity.

Formulas

temperature of maximum crystal growth rate.

How it is measured or obtained

From growth rates measured at several temperatures; for the 31-drug set the way the maximum was obtained is not stated trasi2014a X3; for vemurafenib it is the peak of the measured bell-shaped curve lu2016 R12.

Values in the library
materialvalue or classconditionspaper ids
nimesulide; flutamide; acetaminophen216; 104; 100 µm/sbulk melttrasi2014a R1.a, R1.b, R1.c
felodipine; indomethacin0.86; 0.47 µm/sbulk melttrasi2014a R1.x, R1.z
clotrimazole0.02 µm/sbulk melttrasi2014a R1.ae
vemurafenib α-needles≈106 µm/s at 250 °C; preliminary 106 ± 4 µm/ssupercooled liquidlu2016 R12.o, R25
indomethacin γ; α4.07 × 10⁻⁷; 4.34 × 10⁻⁶ m/spure melttian2017 R7
Pitfalls and disagreements

Several MCVs of the 31-drug set lie far above the highest plotted rate (nimesulide 216 against ≈8.6 µm/s) or below plotted rates (felodipine, indomethacin) trasi2014a X3; the printed limit of the fast group is broken by its own members trasi2014a X2.

← Concepts and formulas

concept · as used in Crystallization tendency

melt viscosity

The steady-shear viscosity of a drug melt at Tm (ηmelt) and below it in the undercooled range; its rise on cooling is expressed as the ratio \eta/\eta_{melt} baird2012 P1, E7. It stands for the molecular mobility that nucleation and growth need baird2012 C4.

Findings

F5, F12.

Related concepts

measures molecular mobility for crystal growth rate and nucleation; used to obtain fragility; related to Tm − Tg temperature gap and glass-forming ability.

Formulas

Vogel-Tammann-Fulcher equation, linear fit of melt viscosity on Tm − Tg.

How it is measured or obtained

Parallel-plate rheometry at 30 s⁻¹, stepwise from Tm downward with remelting between steps baird2012 P1; fitted with the VTF equation baird2012 P5; in drug–polymer melts by the same method tian2017 R13.

Values in the library
materialvalue or classconditionspaper ids
benzamide2.86 × 10⁻³ Pa·s, the lowestat Tmbaird2012 R1.b
ritonavir1.99 Pa·s, the highestat Tmbaird2012 R1.ah
felodipine; nifedipine154.0 and 36.20 × 10⁻³ Pa·sat Tmbaird2012 R1.u, R1.q
class I-A; class III averages≈0.005 and ≈0.11 Pa·s at Tm; ≈0.014 and ≈0.53 Pa·s at 20 °C undercoolingclass averagesbaird2012 R8
17 drugs\eta(T_m) falls linearly with Tm − Tg, R^2 = 0.7368data of baird2012trasi2014a R9
indomethacin with 10% PVPhigher viscosity than with HPMC or HPMCAS at 80–105 °C; pure indomethacin crystallized during measurementrheometrytian2017 R13, C19
indomethacin in a polymermatrix viscosity falls quickly as the drug plasticizes the polymerhot melt extrusionmoseson2018 C45
Pitfalls and disagreements

Single runs at one shear rate; shear itself promoted nucleation at the plate baird2012 X12, C8. The link to crystallization is an association across classes, not shown within classes baird2012 X15. Viscosity does not fully describe felodipine mobility in PVP dispersions kestur2010b C24.

← Concepts and formulas

concept · as used in Crystallization tendency

melting point depression

The lowering of the melting temperature of a crystalline component by the second component of a miscible mixture. Marsac and co-workers describe it as a manifestation of the reduced thermodynamic activity of the drug in the polymer: the dispersed drug reaches the chemical potential of the crystal at a lower temperature marsac2006b C39. In drug–PEG dispersions both branches of the liquidus are depressed, the drug's by dissolving in molten PEG and PEG's by dissolved drug, until they meet at the eutectic [baird2011 E1, E2]. Whether a pair shows depression separates miscible from immiscible pairs marsac2009 C12. New aspects (Miscibility and drug-polymer interactions): For drugs in amorphous polymers the depression is the visible sign of a miscible pair and the route to χ: pairs reported miscible lower the melting point of the drug, immiscible or partially miscible pairs show little or none, and a depression beyond the entropy-only (athermal) prediction means negative χ marsac2009 C5, C12. The offset of the melting endotherm is used because it gives the melting point at the final composition if mixing is complete marsac2009 C16. New aspect (ASD process): for an amorphous polymer the depressed melting point at a fixed composition is called the critical temperature Tc (the temperature at which the last drug crystal has dissolved in the polymer); it is the crystalline solubility temperature of that composition and the lowest product melt temperature at which a crystal-free extrudate is possible in theory, but it is not an upper or lower critical solution temperature moseson2018 P2, C43. A negative Flory–Huggins \chi adds depression beyond the combinatorial entropy moseson2018 C9. New aspect (Physical stability): used as the test of whether a few percent of polymer lowers the driving force for crystallization: no depression means no thermodynamic route kestur2010b C25, C26.

Findings

PEG dispersions: F1, F2, F3, F4, F8, F10. Miscibility and drug-polymer interactions: F6, F7, F8, F9, F11. ASD process: F8, F9, F17, F19, F35, F36, F41. Physical stability: F4, F18, F22, F28, F29.

Related concepts

part of temperature-composition phase diagram; measured by liquidus temperature; depends on enthalpy of mixing and Flory-Huggins interaction parameter; reduces degree of supercooling; related to eutectic; measures Flory-Huggins interaction parameter through the Flory–Huggins relation; depends on drug-polymer miscibility and on the glass transition temperature of the polymer; related to thermodynamic driving force for crystallization. Measured to give formulation critical temperature; sets the lower bound of the HME operating design space.

Formulas

Flory-Huggins melting point depression, Lacoulonche liquidus model.

How it is measured or obtained

By hot-stage polarized light microscopy as the temperature at which the last crystal melts (1 °C/min) and by DSC offset melting (1 °C/min), on fused or physical mixtures that are ground and stored dry [baird2011 P2, P3; zhu2011 P3, P4; zhu2012 P9, P10]. For amorphous polymers the offset is extrapolated to 0 °C/min moseson2018 R5. An interaction parameter follows from the slope of the Flory–Huggins relation at 50–80% drug zhu2011 P5. In the PVP studies: physical mixtures of sieved crystalline drug and polymer (45–75 µm or 45–90 µm), dried for a week over phosphorus pentoxide or nitrogen, geometric mixing, modulated DSC at 1 °C/min under nitrogen, onset (marsac2006b) or offset (marsac2009) of the melting endotherm marsac2006b P1; marsac2009 P1. Only polymers that are liquid-like at the drug melting point work: PVP K12 (Tg 100 °C) gave depression, PVP K29/30 (Tg 168 °C) at 10 °C/min essentially none marsac2006b C29, R11, M3, M4. The fit uses data over 0–0.25 polymer fraction marsac2006b R8. In the extrusion work: cryomilled physical mixtures at 40–90% drug loading (DL), triplicate preparations, 3–5 mg scanned from 25 to 180 °C at six heating rates (0.5, 1, 2, 3, 5, 10 °C/min); the melting point offset temperature is fitted with a second-order polynomial against heating rate and extrapolated to 0 °C/min to give Tc moseson2018 P1, P2, E7. The offset rises with heating rate, for 50/50 IDM:PVPVA from about 133.2 °C at 0.5 °C/min to about 146.1 °C at 10 °C/min moseson2018 R3.f. In the felodipine growth work: DSC offset of the bulk melting endotherm at 1 °C/min of crystalline felodipine cryomilled with 3% PVP K-12; not possible for higher PVP grades, whose Tg lies above the drug melting point kestur2010b P6.

Values in the library
materialvalue or classconditionspaper ids
benzocaine in PEG 3350melting around 89 °C neat, eutectic around 48 °C; text of the other paper: 92 °C neat and about 55 °C for the 20/80 blendfused dispersionszhu2012 C32, M3; zhu2011 R1
ibuprofen; fenofibrate in PEG 3350about 30 °C; about 20 °C by the text, about 24 °C by the ploteutectic depressionzhu2012 C34, X11, R24
chlorpropamide, aceclofenac in PEG 3350liquidus at 40 wt% drug about 55–56 and 86–88 °C against about 98 and 135.5 °C predictedDSC and hot stage[baird2011 R5.e, R6.e]
nifedipine, felodipine in PVP K12decreased linearly over 0–25 wt% polymer; \chi −3.8 and −4.21 °C/min, small particlesmarsac2006b R7, R9
indomethacin in PVP K12\chi −6.16 (onset), −1.94 (midpoint), −0.82 (offset)1 °C/minmarsac2009 R6
indomethacin in PVPVAabout 30 °C at 50% drug loading; 1–4 °C below 30% polymerDSC offsetsmoseson2018 R6
probucol in PEOmelting point 101 °C, Form II only, "may be due to" depressioncompression-moulded dispersion[broman2001 R4.f, C20]
nifedipine in PVP K12onset ≈171 → ≈156.5 °C; ≈144.5 °C at Φ ≈ 0.3831 °C/minmarsac2006b R6
felodipine in PVP K12onset ≈141 → ≈122 °C; ≈115.5 °C at Φ ≈ 0.3831 °C/minmarsac2006b R6
indomethacin in PVP K12offset ≈161.5 → ≈155 °C; onset ≈160 → ≈135.5 °CΦ ≈ 0.23marsac2009 R2
ketoconazole in PVP K12offset ≈150.0 → ≈147.5 °CΦ ≈ 0.23marsac2009 R3
itraconazole with Eudragit E100offset ≈168.0 → ≈167.4 °Cpolymer fraction up to ≈0.54marsac2009 R4
felodipine with 3% PVP, PVP/VA, HPMCAS, PVAc140.7, 142.9, 141.4, 142.6 °C against 141.5 °C: noneoffset, 1 °C/minkestur2010a R1
indomethacin (IDM) with PVPVA, 90/10 to 40/60Tc 159.9 ± 0.3, 157.2 ± 0.4, 151 ± 3, 141 ± 3, 132.3 ± 0.8, 129 ± 6 °C; IDM 160.64 °C; depressions 0.7, 3.4, 9.6, 19.6, 28.3, 31.6 °Coffset extrapolated to 0 °C/minmoseson2018 R5, R6
probucol with PAA (solvent / moulded)melting 110 / 116 °C against 116 °C for pure Form II; read as depression "can be explained by"DSC 20 °C/min for dispersions, 10 °C/min for references[broman2001 R4.c, R4.d, C17, X5]
polymers and surfactants with celecoxib (10% additive)3–4 °C with one additive; 12 °C for TPGS–PVPDSCmosqueragiraldo2014 C18
IDM with copovidone (1:5)solubility temperature about 78 °C (cited)phase diagram[moseson2019 R21]
bicalutamide with PVPVA, 30–90% BCLTc 145 °C at 30% BCL to about 192.3 °C at 90%; drug about 193.8 °Coffset extrapolated to 0 °C/min[moseson2022c R8, R9]
posaconazole with HPMCAS MF, PCZ fractions 0.4–1.0Tc about 161.7 °C at 0.4 to about 168.6 °C for neat PCZ; calculated 135 / 155 / 165 °C at 10 / 25 / 50% DLsame[shah2024 R1, R4, R5, X7]
felodipine with 3 wt% PVP K-12140.7 vs 141.5 °C, within the ±1 °C SD1 °C/minkestur2010b R1.e, M1, X11
indomethacin–PVP (cited)χ23 from melting point depression used as input to the ternary sorption modelliterature[rumondor2010a P5]
acetaminophen with 10% w/w polymers166.7–168.6 against 169 °Cphysical mixtures, DSCtrasi2012a R9
flutamide with 10 mol% polymers108–110 °CDSCtrasi2012b R7
six compounds with 1–20% PHB0–2 °C (1–10% PHB), up to 4 °C (20%)DSC of physical mixturessato2016 R7, X11
indomethacin with 10% polymernamed as a possible reason for the steep inhibition above the growth maximum; not measuredargumenttian2017 C36
Pitfalls and disagreements

Onset, midpoint and offset of the melting event give different depressions and different \chi marsac2009 R6. DSC and hot-stage values differ by up to about 16 °C at some compositions [baird2011 X7]. The low-temperature endotherm of the chlorpropamide and aceclofenac dispersions drifts to 39–49 °C in drug-rich traces, which a simple eutectic would not show [baird2011 R4, X8]. Very slow scans and controlled particle size were needed to see substantial depression with PVP K12, and with PVP K29/32 at 10 °C/min there was essentially none marsac2006b C30, R11. For PEG, the depression of the drug is the cited reason for a smaller driving force and delayed crystallization of benzocaine, but the supercooling in the dispersion was not measured zhu2012 C32, C33. The felodipine premix test read as slow mixing has an unverified direction marsac2009 X6, R8. The linear plot has a small non-zero intercept and the fit range is stated two ways marsac2006b X4. At 3 wt% polymer no depression is measured, and the melting point used there is that of form I in physical mixtures while form II grows kestur2010a R1, X7. Sucrose degrades on melting marsac2009 C14. A polymer with Tg near the drug melting point can hide the depression marsac2009 C22. In the extrusion work, mixtures with little polymer show only 1–4 °C of depression and the extent depends on interaction strength moseson2018 C16, C42. The 40/60 mixture scattered by about ±6 °C and lay above the model line, attributed to viscosity, and was left out of the fit moseson2018 R5.g, C17. In the probucol study depressions of a few °C compare dispersion melts at 20 °C/min with reference melts at 10 °C/min, and the table does not say whether values are onsets or peaks [broman2001 X4, X5]. The same low melting of probucol in moulded PEO is read as depression "may be due to" [broman2001 C20]. The depression of bicalutamide at 30% loading is large (about 49 °C from 193.8 °C to 145 °C, arithmetic on the cited values); that of posaconazole measured at 40% PCZ is about 7 °C (168.6 against 161.7 °C), and the values at 10–25% loading are extrapolations from the fitted line [moseson2022c R8, R9; shah2024 R1, X7]. The felodipine growth test used one composition and one grade only; the conclusion that the driving force is unchanged rests on a 0.8 °C difference inside the error kestur2010b X11. The celecoxib text gives 3–4 °C for one additive while the table gives 1.68–4.49 °C mosqueragiraldo2014 X3, and the depression does not order the celecoxib ternaries mosqueragiraldo2014 X13.

← Concepts and formulas

concept · as used in Crystallization tendency

metastable zone

Evidence table: metastable zone. Content from the ASD dissolution chapter (2008, 2010) and the LLPS and Bioperformance chapters.

The range of supersaturation in which a solution is supersaturated with respect to the crystal but does not nucleate spontaneously, so that existing crystals can grow while new ones hardly form; above it lies the labile zone of spontaneous nucleation [konno2008a C27]. In the scheme of two crystallization routes, nucleation and growth deplete the solution once its concentration exceeds the metastable zone alonzo2010 C43.

Findings

F5.

Related concepts

related to nucleation, classical nucleation theory, supersaturation and binodal.

Formulas

classical nucleation theory.

How it is measured or obtained

Not measured in the ASD dissolution papers; inferred from the shape of concentration–time curves [konno2008a C27, X4]. On a temperature–concentration diagram of ritonavir the binodal and spinodal of liquid–liquid separation were mapped against the crystal solubility ilevbare2013a R11.

Values in the library
materialvalue or classconditionspaper ids
felodipine with HPMC or HPMCAS after an initial droptaken to sit in the metastable zone ("it appears")pH 6.8, 37 °C[konno2008a C27, R12]
ritonavir (LLPS chapter)crystal-supersaturated region below the binodal; binodal–spinodal gap smallwaterilevbare2013a C32, C34
felodipine, nifedipine (Bioperformance)the amorphous solubility lies within the metastable zone for crystal nucleationbufferraina2014a C30
Pitfalls and disagreements

The metastable-zone limit of felodipine was not measured, so the reading of the plateaus as growth-limited rests on argument [konno2008a C27]. Seeded and unseeded tests later located the effect of HPMC on felodipine mainly in nucleation [alonzo2012 R17].

← Concepts and formulas

concept · as used in Crystallization tendency

moisture-induced crystallization

Crystallization of an amorphous drug, alone or in a dispersion, that sorbed water starts or speeds up. In the library it is measured as a nucleation rate, a crystal growth rate or a crystallized fraction that rises with storage relative humidity [konno2008 C11; rumondor2010 C8, C9; rumondor2009b C7]. Two routes are distinguished: water raises molecular mobility (plasticization) and may raise the drug's chemical potential, or water first splits a dispersion into drug-rich and polymer-rich amorphous phases, from which the drug-rich phase crystallizes [marsac2008a C1, C26; rumondor2009c C5; marsac2010 C25]. New aspect (Supersaturation from ASD dissolution): it serves as the solid-state comparison for the polymer ranking in dissolution: dry felodipine dispersions with PVP, HPMC and HPMCAS nucleated alike, whereas with moisture HPMCAS and HPMC inhibited crystallization better than PVP (cited) [konno2008a R18]. The resveratrol dispersions that resisted crystallization at high humidity (PVP, E100) also stayed amorphous in buffer but gave no concentration gain, and those that crystallized at high humidity (HPMC, HPMCAS, CMCAB) also crystallized in buffer at 35% drug wegiel2015 R21, C7, C9. Indomethacin dispersions at 60% drug with HPMC or EPO crystallized within 5 days at 40 °C/75% RH, the EPO–HPMC ternary not within 100 days [xie2017 R26].

Findings

Physical stability: F9, F11, F15, F24, F25, F33, F34. Supersaturation from ASD dissolution: F4, F6, F16, F20.

Related concepts

depends on plasticization by water and water sorption; can follow moisture-induced phase separation; part of solid-state crystallization of amorphous drugs; related to hygroscopicity; related to physical stability and crystallization inhibition by polymers.

Formulas

none specific; the water uptake that drives it is described by Flory-Huggins water sorption isotherm.

How it is measured or obtained

Storage over saturated salt solutions at fixed RH and counting of birefringent nucleation sites in spin-coated films konno2008 P3, P4; growth of existing crystals in films at fixed RH rumondor2010 P3, P4, P5; PXRD crystallinity of stored powders with PLS calibration [rumondor2009b P5]; AFM and polarized light on films after 1–16 h at 94% RH marsac2010 P14, R19. Outside the subcategory, SHG imaging of spray-dried dispersions at 40 °C and 75–82% RH correasoto2017 R4, C15.

Values in the library
materialvalue or classconditionspaper ids
amorphous felodipine filmnucleation ≈1.4 × 10⁵ → 2.9 × 10⁶ m⁻³ s⁻¹0 → 75% RH, 22 °Ckonno2008 R1.a, R3.a
amorphous nifedipine filmnucleation ≈4.7 × 10⁵ → 6.8 × 10⁶ m⁻³ s⁻¹0 → 75% RHmarsac2008a R1.a
amorphous felodipine filmgrowth ≈0.030 → 0.066 mm/min (13–80% RH), 2.3–2.8 mm/min at 87–93% RH, with needleshumidity stage, 25 °Crumondor2010 R1, C9
felodipine–PVP 25% powder1.2% (0% RH) to 82.1% (93% RH) crystallized after ≈494 days22–25 °C[rumondor2009b R9.a, R9.g]
felodipine–HPMCAS 50% powder9.2–16.6% at every RH after ≈490 days22–25 °C[rumondor2009b R9]
felodipine–PVP 50% filmcrystals by 4 h at 94% RH, after phase separationAFM, polarized lightmarsac2010 R19, C24
FTM–HPMCAS spray-dried dispersion (outside)faster crystallization at 82% than at 75% RH40 °C, SHGcorreasoto2017 C15
Accord tacrolimus capsules (outside)rapid crystallization in the powder blend40 °C/75% RH[purohit2019 C33]
resveratrol with HPMC, HPMCAS, CMCAB, PAAcrystalline at the first sampling at 84% RH, then fully; PVP and E100 not25 °Cwegiel2013b R12, R14, R16
curcumin dispersionsat 84% RH: E100 > 350, cellulosics 75, PVP 22, neat 14 days25 °Cwegiel2014 R13
indoprofen–PVPseparation at 54% RH, crystals from 75% (30% PVP) or 84% RH (50% PVP)22 °C films[rumondor2011 R11, R14]
lumefantrine–CAP 40:60 precipitated into water (9% water)crystalline from the first day at 40 °C/75% RH; amorphous 3 months under vacuumPXRD[bhujbal2021 R4.a, R5, R10, C17]
tacrolimus generic capsule contents≈36, 68 and 84.5% at 30, 35 and 40 °C/75% RH, ≈11% at 40 °C/54% RH, 4 weeksopen dishtrasi2017 R14
felodipine dispersions with PVP, HPMC, HPMCAS (cited)no difference dry; HPMCAS ≈ HPMC > PVP at high RHstorage[konno2008a R18]
flutamide and ezetimibe HPMCAS dispersions (Solid-state methods)first storage time with detected crystallinity, by methodstoragecorreasoto2017 R1
tacrolimus capsules (Bioperformance)% crystallized against storage timestorage[purohit2019 R1]
resveratrol dispersions (storage, cited)PVP, E100 stable; HPMC, HPMCAS, CMCAB crystallized at high RHstoragewegiel2015 R21
indomethacin 60% with HPMC; EPO; EPO–HPMCcrystals within 5 days; within 5 days; amorphous beyond 100 days40 °C/75% RH[xie2017 R26]

Evidence table: moisture-induced crystallization.

Pitfalls and disagreements

The growth jump of felodipine above 80% RH has no counterpart in water uptake or Tg, and its cause is "currently unknown" rumondor2010 C31, C33. Bulk powder of pure amorphous felodipine crystallized within 12 days at 0% as well as at 93% RH, so at that resolution water made no difference [rumondor2009b R12, X8]. Whether crystallization follows the wet one-phase route or the phase-separated route changes how polymer content acts (no effect of PVP level at 93% RH) [rumondor2009b C20, C21]. For resveratrol the authors "do not fully understand" why humidity acted more than heat, and the contrast fails at 120 °C wegiel2013b C40, X8. In the precipitated lumefantrine series water content varies with the polymer, so the water effect is inferred [bhujbal2021 C51, C56]. A solid-state ranking under moisture need not hold in dissolution: HPMCAS and HPMC were alike in humid solids but HPMCAS dispersions gave far more supersaturation, and the authors suggest the inhibition mechanisms "may vary" with the amount of water [konno2008a C33, C34].

← Concepts and formulas

concept · as used in Crystallization tendency

molecular recognition

The specific fit between molecules that lets them assemble in a given arrangement; in crystal engineering hydrogen bonding is the most reliable directional interaction for it [vaneerdenbrugh2011a C2]. In the physical-stability papers it is the name for how an additive may order the drug locally or enter its pre-nuclei, an effect that classical nucleation theory does not directly contain trasi2012a C48, C49.

Findings

F17.

Related concepts

part of nucleation; contrasts with classical nucleation theory; related to hydrogen-bond donor and acceptor groups and self-association.

Formulas

classical nucleation theory (which does not contain it) trasi2012a C49.

How it is measured or obtained

Not measured directly in these papers; it is inferred when an additive changes nucleation in a way that growth inhibition, Tg or hydrogen bonding do not explain trasi2012a C50, C52; trasi2012b C47.

Values in the library
materialvalue or classconditionspaper ids
acetaminophen with 10% w/w PAA26 nuclei per cycle against 10.8; PAA "may" exert a local ordering effecthot-stage cycle to −30 °Ctrasi2012a R12.e, C50
flutamide with 10 mol% nilutamide (structural analogue)one nucleus in the imaged region; nilutamide "may" enter prenuclei and block themfilm cooled to −30 °C and reheatedtrasi2012b R12, C47
15 drugs with 7 polymers (other chapter)hydrogen-bond recognition rules applied to choose polymers that prevent crystallizationspin-coated films[vaneerdenbrugh2011a C51]
Pitfalls and disagreements

Both physical-stability uses are hypotheses: local ordering of acetaminophen near PAA and nilutamide inside flutamide prenuclei were not observed trasi2012a C50; trasi2012b C47. The acetaminophen counts behind the PAA reading add up nucleation over zones of different width, so PAA need not raise the rate per unit time trasi2012a X8.

← Concepts and formulas

concept · as used in Crystallization tendency

monotropy

A polymorph pair in which one form is the more stable at every temperature up to melting, so the other can only convert into it; the free-energy curves would cross only at a virtual transition temperature above both melting points lu2016 C51; trasi2014b C11.

Findings

F7.

Related concepts

tested by Burger-Ramberger rules; part of polymorphism.

Formulas

Burger-Ramberger heat-of-fusion rule, Hoffman equation.

How it is measured or obtained

By the heat-of-fusion rule from melting data lu2016 C50; trasi2014b C11; by Hoffman free-energy curves of both forms lu2016 R22; supported by the direction of observed conversions lu2016 C38.

Values in the library
materialvalue or classconditionspaper ids
vemurafenib α and βmonotropic; virtual transition 319 °Cheat-of-fusion rule and Hoffman curveslu2016 C50, C51, R22
nilutamide stable form and form Imonotropic; form I about 3.4 kJ/mol less stable at 25 °Cmelting datatrasi2014b C11, R7
Pitfalls and disagreements

The vemurafenib virtual transition temperature depends on which α heat of fusion is used lu2016 X1; the room-temperature conversion that supports α as the stable form is not shown lu2016 X13.

← Concepts and formulas

concept · as used in Crystallization tendency

nucleation and crystal growth

The two steps of crystallization, the formation of new crystals and the growth of existing ones; they have different temperature windows and can respond differently to the same additive, and overall crystallization of a powder is nucleation followed by growth trasi2012a C35, C53; kestur2013b C35, C47. The term is also used for a route of amorphous–amorphous phase separation, in which discrete domains of the new phase nucleate and grow, as opposed to spinodal decomposition [saboo2017 C40; yang2022b C50, C55].

Findings

F17, F21, F26, F31, F32, F33.

Related concepts

part of solid-state crystallization of amorphous drugs; consists of nucleation and crystal growth rate; located by nucleation temperature zone.

Formulas

classical nucleation theory, linear crystal growth rate from size-time slope.

How it is measured or obtained

Separated by growing seeded or pre-formed crystals on a hot stage and by locating nucleation zones and counting nuclei in cooling–reheating cycles trasi2012a P2, P6, P7; trasi2012b P2, P6, P7; or inferred by comparing overall powder crystallization with linear growth rates kestur2013b C47.

Values in the library
materialvalue or classconditionspaper ids
acetaminophennucleation zone 50–60 °C, where growth is ≈0.04–0.23 µm/s against ≈45 µm/s at 100 °CDSC and hot stagetrasi2012a R12.a, R8.a, C39
acetaminophen with PAA or HPMCASPAA slows growth most but gives most nuclei; HPMCAS the reverse10% w/wtrasi2012a C46
flutamide with PVP or nilutamidePVP slows growth about tenfold without a nucleation effect; nilutamide nearly stops nucleation10 mol%trasi2012b C7, R11.e, R12
felodipine powders with 3% PVPearly crystallization 60–77-fold slower; surface and bulk growth ≈2- and ≈7-fold30 °Ckestur2013b R15, R12, C34
miconazole–PVPVA films after humid casting or storagediscrete drug-rich domains in a polymer-rich matrix, read as separation by nucleation and growth ("may occur")confocal, AFM[saboo2017 C40]
ritonavir–PVPVA films with SDS or Tween 80 vs binary and Spansnucleation and growth vs spinodal decomposition, read from the first morphologyconfocal[yang2022b C55]
tacrolimus genericfall in concentration after 40–50 min read as growth on seeds present before storage ("some evidence")dissolution, 100 mLtrasi2017 C44
Pitfalls and disagreements

Nucleation was counted per thermal cycle or inferred from DSC peak temperatures and rate ratios, never measured as a rate at storage temperature trasi2012a X8; trasi2012b X4; kestur2013b X7.

← Concepts and formulas

concept · as used in Crystallization tendency

nucleation induction time

The time from creating a supersaturation until crystals are first detected, t_{ind} = t_n + t_g: the time to form stable nuclei plus the time for them to grow to a detectable size; if growth is quick, t_{ind} \propto J^{-1} and a longer time means a slower nucleation rate ilevbare2013c E2, E3, C14. It depends on the detection method, since crystals are seen earlier by SHG than by a probe ratio or light scattering [jackson2014 R25, R17].

Findings

F3, F4, F5, F6, F13, F14, F15, F25, F27, F28, F29, F32, F34, F35, F37, F38, F39, F40, F41, F42, F43, F44.

Related concepts

part of nucleation; gives nucleation rate; modified by nucleation inhibition; measured on supersaturation.

Formulas

induction time decomposition, relative nucleation rate from seeded and unseeded induction times, classical nucleation theory.

How it is measured or obtained

A methanol stock of drug is pumped or injected into stirred buffer with or without predissolved polymer and the signal is followed: extinction at a wavelength where the drug does not absorb (280 or 350 nm) or the fall in UV absorbance, taking the crossing of regression lines through the flat and rising (or falling) parts as the time; pyrene fluorescence and SHG as second read-outs; triplicate [alonzo2012 P5; ilevbare2013c P9, P10; jackson2014 P8]. The stated supersaturation uses a methanol-free solubility; the methanol fraction is about 0.5% v/v ilevbare2013c X16.

Values in the library
materialvalue or classconditionspaper ids
celecoxib; efavirenz; ritonavir, no polymerabout 2.4; 5.1; 120 minat the calculated amorphous solubility, 37 °Cilevbare2013c R8, R9.a
ritonavir, CAB Adp 0.81; efavirenz, CAB Adp 0.81; celecoxib, HPMCabout 244; 28.0; 12.6 min5 µg/mL polymerilevbare2013c R9
celecoxib at 11 µg/mL, none against HPMC27.5 ± 0.7 against 855.5 ± 6.0 minS 7.3ilevbare2013c R15
felodipine, none against 1 µg/mL HPMC15.3 against 136.5 minS 10, 25 °C[alonzo2012 R16]
danazol, none; PVP; HPMC; HPMC-AS6–10; 88–162; 111–285; 250–516 min10–20 µg/mL, 25 °C[jackson2014 R17]
celecoxib 22 µg/mL, 5 µg/mL HPMCAS (ASD dissolution chapter)held for 16 h; J_0/J_p about 4,00037 °C[xie2016a R3, R13]
paclitaxel 30.7 µg/mL (S 88) (ASD dissolution chapter)within 6 min; more than 16 h with 20 µg/mL HPMCAS37 °C[que2018 R3]
indomethacin S 6 (ASD process chapter)about 40 min; more than 12 h with 5–50 µg/mL PVPVApH 4.5, 37 °C[moseson2020a R3]
11 drugs, 0.1% STC and SDS, matched C/Ceqbuffer 7–87 min; STC 15 to >1000; SDS shorter for 7 of 925 °C, telaprevir 37 °Cchen2015 R3
acetaminophen, S 1.595% of vials 26 min; first crystals 150–785 s20 °Ctrasi2015 R1, R10
celecoxib, nevirapine, flibanserin, 13 bile salts3 min to >16 h (celecoxib); 0–577 (nevirapine); 0–623 min (flibanserin)pH 6.5, 37 °C[li2016a R6]
telaprevir, 80 and 150 µg/mL, 26 polymersHPMC >700 and 243; none 14–15 minpH 6.8, 37 °C[mosqueragiraldo2016 R4]
telaprevir δ 11.5 and 28.8, six bile salts30 min and immediate in buffer; >960 min with 1.86 mMpH 6.5, 37 °Clu2017 R3, R4
danazol 15 µg/mL, PVP, HPMC, HPMC-AS 1–1000 µg/mLabout 5 min; 26–412 min25 °C[jackson2016 R17]
atazanavir without additive, five protocolsabout 40, 92–102, 170, 200, 253 ± 65 minnear the amorphous solubility, 37 °C[mosqueragiraldo2018a R1; elkhabaz2021 R6; indulkar2018 R19; indulkar2019b R3; enright2020 R2]
posaconazole without additive, three protocolsabout 5–8, 8.4, 29 ± 5 minpH 6.5, 37 °C[elkhabaz2021 R7; duong2022b R10; shah2025 R3]
enzalutamide without polymerabout 5–15 min, 30–120 µg/mLpH 6.5wilson2021 R1
Pitfalls and disagreements

The assumption t_n \gg t_g was not tested; strong inhibitors give gradual onsets whose fitted times depend on where the lines are drawn (rise above baseline at about 25 min against a fitted 31.5 min; about 180 against 260 min) ilevbare2013c C14, X14. Methods differ by up to about 40% for single conditions [jackson2014 X5]. The same polymer, drug and concentration gave 10–13 min in one laboratory and more than 8–16 h in another [ilevbare2013c R9; xie2015 R1; xie2016a R3]. The efavirenz run was at S about 2.3, not 15 ilevbare2013c X4. Replicate scatter is below 15% ilevbare2013c R17. Values above 960 or 720 min are lower bounds set by the run length lu2017 X16; chen2015 X7. The control induction time of one drug differs up to about sixfold between protocols, so effects compare only as ratios to a control in the same protocol [mosqueragiraldo2018a R1.b; enright2020 R2]. Several papers do not identify the scattering particles at the induction time [indulkar2018 X14; enright2020 X15].

← Concepts and formulas

concept · as used in Crystallization tendency

nucleation inhibition

The slowing of crystal nucleation from a supersaturated solution by an additive, measured as a longer nucleation induction time or a lower nucleation rate. Ilevbare and co-workers propose that a polymer inhibits nucleation by joining clusters of solute molecules and hindering their reorganization into an ordered nucleus, which needs a polymer with more affinity for the solute than for water or for itself ("we assume", "would be expected") ilevbare2013c C4, C5, C35. How additives slow nucleation is more speculative than how they slow growth, because the molecular events cannot be observed ilevbare2013c C9. New aspect (Supersaturation from ASD dissolution): for celecoxib released from dispersions, the authors take the polymer effect on nucleation as "of paramount importance" to the desupersaturation profile, because growth needs nuclei xie2015 C40. Extended induction times with polymers indicate a decreased nucleation rate, PVP-K12 being the least effective and HPMCAS or HPMC showing no discernible nucleation or growth for 8 h xie2015 C42, R1; the greater effect of the cellulose derivatives "may relate" to a greater ability to interact with a dense liquid cluster of celecoxib than the more hydrophilic PVP, which also interacts with the solvent xie2015 C46. With 5 µg/mL HPMCAS the relative effect on nucleation (~4000) was far larger than on growth (70) ("it can be surmised") [xie2016a C40, R13].

Findings

Crystallization from solution: F6, F13, F14, F15, F20, F27, F28, F31, F32, F33, F36, F40. Supersaturation from ASD dissolution: F4, F5, F20, F29.

Related concepts

contrasts with crystal growth inhibition; part of crystallization inhibition by polymers; measured by nucleation induction time and nucleation rate; related to polymer hydrophobicity; related to interfacial tension.

Formulas

relative nucleation rate from seeded and unseeded induction times, induction time decomposition, classical nucleation theory.

How it is measured or obtained

Unseeded induction times with and without polymer under identical conditions (see nucleation induction time), optionally combined with seeded growth rates into the nucleation-rate ratio J/J_0 [alonzo2012 P10, E10]. In the celecoxib dispersion papers: solvent shift to 22 µg/mL celecoxib in pH 6.8 phosphate buffer at 37 °C with 5 µg/mL dissolved polymer, the onset read from a rise in 350 nm extinction with a fall in 249 nm absorbance xie2015 P4, R1.

Values in the library
materialvalue or classconditionspaper ids
felodipine, 1 µg/mL HPMCJ/J_0 7.05 × 10⁻⁴ ("more than a thousand fold"); induction 15.3 → 136.5 minS 10, 25 °C[alonzo2012 R16, R17, C38]
ritonavir; efavirenz; celecoxib, best polymer120 → 244; 5.1 → 28.0; 2.4 → 12.6 min5 µg/mLilevbare2013c R9, R12
celecoxib at S 7.3, HPMC27.5 → 855.5 min (31-fold)5 µg/mLilevbare2013c R15
celecoxib, 5 µg/mL HPMCAS (ASD dissolution chapter)J_0/J_p about 4,00022 µg/mL drug[xie2016a R13]
paclitaxel, HPMCAS (ASD dissolution chapter)"highly effective" at S 8820 µg/mL polymer[que2018 R3, C13]
indomethacin, PVPVA 5–50 µg/mL (ASD process chapter)"highly effective" at S 6; stays supersaturated more than 12 hpH 4.5[moseson2020a R3, C11]
11 drugs, 0.1% STCinduction time lengthened for all, eight by more than 10-fold25 °Cchen2015 R3, X6
telaprevir, 1.86 mM STC, SGCrelative nucleation rate J/J₀ 2.5 × 10⁻³ and 2.9 × 10⁻⁶ (recomputed 2.58 × 10⁻³ and 3.06 × 10⁻⁶)δ 11.5lu2017 R10, X1
acetaminophen, S 1.5J lowered 18-fold (HPMCAS) and 36-fold (HPMC); raised 2.6-fold by PAA10 µg/mLtrasi2015 R3
telaprevir, 1.86 mM binary and ternary bile salt mixtures>960 min against about 30 in buffer; 3.72–5.58 mM mixtures about 255 minδ 11.5, 37 °C[lu2018 R3, R4]
nine drugs, 5 µg/mL A5aatazanavir 40 → >300, telaprevir 15 → 91 minpH 6.8, 37 °C[mosqueragiraldo2018a R1]
celecoxib 22 µg/mL: no polymer; 5 µg/mL PVP-K12; PVP-K29/32; HPMCAS or HPMCwithin 5 min; ≈1 h; ≈2 h; more than 8 hpH 6.8, 37 °Cxie2015 R1
delamanid at 0.76 µg/mL with 300 µg/mL polymermore than 1000 min with PVPVA, HPMCP and HPMCAS; about 50 min with HPMC; 10.2 min with Eudragit L100 against 10.7 min without polymerPBS pH 6.5, 37 °Cduong2022a R4, R2
racemic praziquantel at 1.1 mg/mL with 300 µg/mL polymer10.9 (PVPVA), 14.3 (HPC), 15.7 (PVA), 24.5 (HPMC 603) and 50.5 min (HPMCAS-LF); HPMCAS-MF and HPMC E5 LV beyond the 83-min runPBS pH 6.5, 37 °C[polyzois2024 R3]
R-praziquantel at 1.4 mg/mL37 min (HPMCAS-MF) and 15.5 min (HPMC E5 LV)PBS pH 6.5, 37 °C[polyzois2024 R4]
Pitfalls and disagreements

The mechanism (cluster association, polymer–solute affinity, self-association, chain rigidity) was not measured; the induction time includes growth to a detectable size ilevbare2013c C5, C39, C43, X14. The nucleation-rate ratio of felodipine rests on one pair of induction times entering to the fourth power [alonzo2012 X7]. Rankings of polymers differ from rankings of growth inhibition (F6) ilevbare2013c C40. In the celecoxib dispersion papers the induction time includes growth to a detectable size xie2015 E1; the HPMCAS ratio of ~4000 takes the 960-min end of the run as the induction time, so it underestimates the effect [xie2016a R13, C40]; celecoxib clusters and their interaction with polymers were not measured xie2015 C46. A ranking made with a racemate did not hold for its enantiomer [polyzois2024 R4, C8], and the praziquantel run length capped the two best polymers at 83 min so they cannot be ranked against each other [polyzois2024 R3].

← Concepts and formulas

concept · as used in Crystallization tendency

nucleation rate

The number of stable crystal nuclei formed per unit volume and time, J. Classical nucleation theory gives J = A\exp(-\Delta G_{crit}/kT) with a barrier that falls steeply with supersaturation, \Delta G^{*} \propto 1/(\ln S)^{2}; where the induction time is dominated by nucleation, t_{ind} \propto J^{-1} ilevbare2013c E3, E4; hsieh2012 E7, E8. The nucleation-rate ratio with and without a polymer is estimated from seeded and unseeded induction times [alonzo2012 E10]. New aspect (Physical stability): in the solid-state papers of the library it is the slope of the volume density of birefringent sites against time in amorphous films, in m⁻³ s⁻¹ konno2006 C12, R3; marsac2008a E4; in solution papers it is derived from induction times or counts of particles or SHG signals per time [alonzo2012 E10; elkhabaz2021 E5].

Findings

Crystallization from solution: F6, F14, F27, F32, F33, F34, F42. Physical stability: F1, F2, F9, F10, F11.

Related concepts

inverse of nucleation induction time; modified by nucleation inhibition; described by classical nucleation theory; measures nucleation; set by kinetic barrier to nucleation and thermodynamic driving force for crystallization; related to crystal growth rate.

Formulas

relative nucleation rate from seeded and unseeded induction times, classical nucleation theory, induction time decomposition, depth of field of a microscope objective, Arrhenius equation.

How it is measured or obtained

Not measured as an absolute rate in the papers of the crystallization-from-solution chapter. The ratio is J/J_0 = (t_s/t_{s,0})^{3}(t_{u,0}/t_u)^{4}, with the seeded induction-time ratio replaced by the growth-rate ratio R/R_0 at the same supersaturation and polymer level [alonzo2012 P10, E9, E10]. In the ASD dissolution chapter the same relation was applied to celecoxib [xie2016a R13]. In the solid-state papers: spin-coated films stored at a fixed RH and temperature; sites counted under polarized light in 12 areas per film, converted from area to volume with the depth of field of the objective (0.0144 mm at 10×), triplicate, film edges excluded konno2006 P2, E1; marsac2006a P4; konno2008 P4. Sites are counted once they reach about 1 µm konno2008 C38. In SHG images of griseofulvin the number density of crystal sites was followed over time with a 60 µm depth of field [wanapun2010 R17].

Values in the library
materialvalue or classconditionspaper ids
felodipine, 1 µg/mL HPMCJ/J_0 7.05 × 10⁻⁴ (text: more than thousand-fold; conclusions: factor of 1000)S 10, 25 °C[alonzo2012 R17, C38, C46]
celecoxib, 5 µg/mL HPMCASJ_0/J_p about 4,000 (unseeded time taken as 960 min)22 µg/mL drug, 37 °C[xie2016a R13]
acetaminophen, S 1.5J 1465 (none), 3820 (PAA), 81.86 (HPMCAS), 40.68 (HPMC) # m⁻³ s⁻¹1.2 mL vials, 20 °Ctrasi2015 R3
telaprevir with STC, SGCJ/J₀ 2.5 × 10⁻³, 2.9 × 10⁻⁶δ 11.5lu2017 R10
atazanavir, posaconazole, six mediaSHG count rate about 0.1–1.6 and 0.05–48 counts/min37 °C, relative per field[elkhabaz2021 R14, E5, X13]
posaconazole, PBS and three mediaJ from induction times, fitted for A and γ37 °C, S above about 7–13duong2022b R15, X10
amorphous felodipine≈1.3 × 10⁵ m⁻³ s⁻¹; ≈2–3 × 10³ with 3% PVP, HPMCAS or HPMC0% RH, 22 °Ckonno2006 R4
amorphous nifedipine vs felodipine0.39 vs 0.17 × 10¹⁰ m⁻³ day⁻¹0% RH, 22 °Cmarsac2006a R4.a
nifedipine and felodipine with 7% PVP≈1.4 × 10³ and ≈2.7 × 10² m⁻³ s⁻¹0% RHmarsac2006a R16.b
felodipine with 3% PVP vs HPMCAS≈8.6 × 10⁵ vs ≈1.3 × 10⁴ m⁻³ s⁻¹75% RH, 22 °Ckonno2008 R3.b
nifedipine vs felodipine with 25% PVP≈4.3 × 10⁵ vs ≈2.3 × 10³ m⁻³ s⁻¹75% RHmarsac2008a R1.f, R1.l
felodipine (activation energy)145 kJ/mol below Tg; 201 with 3% PVPArrhenius, ≈25–75 °Cmarsac2006a R3
griseofulvin melt (SHG imaging)number density ≈45 → ≈600 mm⁻³ over ≈62–285 min125 °C, SHG[wanapun2010 R17]
Pitfalls and disagreements

The ratio rests on one pair of induction times without uncertainty and on a seeded ratio standing in for a seeded induction-time ratio; a 10% change in the polymer-case time changes J/J_0 about 1.5-fold [alonzo2012 X7, C11]. The printed inverse of 0.607 is 1.643 against 1.647, with J/J_0 7.05 or 7.06 × 10⁻⁴ [alonzo2012 X6]. The celecoxib value is a lower bound because no desupersaturation occurred within the experiment [xie2016a R13]. In the film papers absolute rates are per depth-of-field volume and comparable only within one counting method; in one paper Table II and two figures disagree about fourfold, and the felodipine-alone rate of that paper's Fig. 9 (≈3 × 10⁴) is about fourfold below the other papers' ≈1.3–1.4 × 10⁵ (synthesis) marsac2006a X1; konno2006 R4.a. The 2006 methods describe the area-to-volume conversion as multiplication where division is meant konno2006 X6. Site density rose roughly linearly, but early points can lie below the fitted line, so a short lag is not excluded konno2006 X8.

← Concepts and formulas

concept · as used in Crystallization tendency

nucleation temperature zone

The temperature range in which nuclei form at a useful rate; it usually lies below the temperature of fastest growth, and storing outside it helps prevent crystallization trasi2014b C3. Each polymorph may have its own zone trasi2014b C38; lu2016 C42.

Findings

Crystallization tendency: F3, F4. Physical stability: F17.

Related concepts

part of nucleation and of nucleation and crystal growth; includes sub-Tg nucleation; contrasts with the growth window of crystal growth rate; related to glass-forming ability.

Formulas

critical cooling rate to avoid nucleation.

How it is measured or obtained

By cooling the melt to successively lower end points and reheating; the highest end point that gives crystallization on reheating bounds the zone trasi2014b P1; trasi2012a R12; by nuclei counts after holds at different temperatures trasi2014b P7. The zone limits depend on the cooling and heating rates and serve only to compare samples trasi2012a C13.

Values in the library
materialvalue or classconditionspaper ids
nilutamidestrong zone near −30 °C; weaker rise from about 50 °C, about 30 °C above TgDSC; surface countstrasi2014b R5, R9, C41
acetaminophen50–60 °CDSCtrasi2012a C22
flutamide10–40 °CDSCtrasi2012b R10
acetaminophen with 10% w/w HPMC, HPMCAS, E100, PVPh, PVPVA, PVP, PAA10–60, 10–45, 40–60, 40–70, 30–60, 30–50, −10–30 °CDSC cool–reheattrasi2012a R12
acetaminophen with 15% w/w PVP, PAA, HPMCASno crystallization in the testsametrasi2012a R12
flutamide with 10 mol% PDMA, PVAc, PVPH, PIPA, PVP10–30, 20–40, 30–40, 20–30, 20–40 °Csametrasi2012b R10
vemurafenib αapproximately 170–270 °C (inferred)hot stagelu2016 C43
indomethacin γ; α (cited)maxima at about Tg and about 20 °C higherliteraturetrasi2014b R32
Pitfalls and disagreements

The method passes the zone on both cooling and reheating, so the two contributions are not separated trasi2012a X9; the polymorph of the counted nuclei was not identified for nilutamide trasi2014b X6. Storage inside the zone gives high nucleation; a polymer that lowers the zone (HPMCAS for acetaminophen) "could potentially" still give more nuclei at room temperature than another polymer trasi2012a C47.

← Concepts and formulas

concept · as used in Crystallization tendency

nucleation

Nucleation is the formation of crystal nuclei, the first of two dependent steps of crystallization, each with its own rate and temperature dependence baird2013 C32; kestur2010a C27. In classical nucleation theory the barrier to a critical nucleus grows with the cube of the interfacial free energy and falls with the square of the driving force vaneerdenbrugh2010a E3; the library adds molecular recognition as a further factor trasi2012a C48; konno2006 C33. This draft comes from the crystallization-tendency chapter (supercooled melts and glasses); nucleation from solution is covered by other chapters. New aspects (Miscibility and drug-polymer interactions): For indomethacin and PVP the proposal is that a critical γ nucleus is built of acid dimers, so that "the formation of a critical nucleus of indomethacin would require the breaking of the PVP-indomethacin hydrogen bond"; the same logic is expected to hold for α, which hydrogen-bonds in chains taylor1997 C43, C44. During solvent removal nucleation and growth can occur once the drug's solubility in the drying solution is exceeded, and a polymer may alter the supersaturation reached vaneerdenbrugh2010b C29. Pure nifedipine and felodipine nucleate much faster than dispersions with even small amounts of polymer, which is the main argument that the polymer acts through kinetics marsac2006b R16, C42. New aspect (Crystallization from solution): nucleation from supersaturated aqueous solution differs from nucleation in a supercooled melt in that the driving force is the supersaturation S and the read-out is the induction time at fixed S; it is "energetically more demanding" than growth, so at some supersaturations crystals grow while new nuclei do not form ilevbare2013c C10, E3. Polymers can slow it several-fold at the supersaturation of amorphous dissolution and by much more at lower supersaturation (F13, F14), and for felodipine far more than they slow growth (F6) [alonzo2012 C46; ilevbare2013c R12, R15]. In a two-phase solution the droplet interface is a candidate site (see heterogeneous nucleation) [jackson2014 C45, C53]. New aspect (Physical stability): the physical-stability papers treat nucleation as the initiating event that a dispersion must suppress, framed by classical nucleation theory with a thermodynamic barrier and a kinetic barrier, while noting that nucleation in films is "most likely heterogeneous" marsac2006a C5; konno2006 C33. New aspect (Supersaturation from ASD dissolution): three readings from dissolving dispersions: a fast initial drop of a highly supersaturated felodipine solution as rapid nucleation, faster and giving more particles at higher concentration [konno2008a C24]; immediate crystallization of drug released at very high local supersaturation from a 10% drug dispersion [alonzo2011 C29]; and nucleation inside drug-rich droplets, favoured by the high drug concentration and an expected low crystal–liquid interfacial tension [ilevbare2013d C39, C40].

Findings

Crystallization tendency: F3, F4, F7, F15, F16, F18, F19, F21. Miscibility and drug-polymer interactions: F1, F9, F10, F15. Crystallization from solution: F3, F6, F13, F14, F20, F34, F36. Physical stability: F1, F2, F9, F10, F16, F17, F20, F29. Supersaturation from ASD dissolution: F3, F5, F13.

Related concepts

part of the process together with crystal growth rate (contrasts with); measured by nucleation temperature zone; modelled by classical nucleation theory; includes heterogeneous nucleation and sub-Tg nucleation; related to quenched-in nuclei and prenucleation clusters; depends on thermodynamic driving force for crystallization; related to crystallization inhibition and kinetic stabilization; includes secondary nucleation; inhibited by nucleation inhibition; quantified as nucleation rate; controlled by kinetic barrier to nucleation; see molecular recognition and nucleation and crystal growth; related to nucleation induction time and metastable zone.

Formulas

classical nucleation barrier, critical cooling rate to avoid nucleation, classical nucleation theory, induction time decomposition, relative nucleation rate from seeded and unseeded induction times, Arrhenius equation, depth of field of a microscope objective.

How it is measured or obtained

Directly, by the number of nuclei grown to visible size after a hold (two-step method) trasi2014b P7 or by nucleation-site density over time in films marsac2006a R1; by locating a nucleation zone with DSC cooling to successive end points and reheating trasi2014b P1; trasi2012a R12; indirectly, from critical heating rates after slow and fast cooling baird2013 C38 or from the absence of crystallization on reheating trasi2014b C39. In solution, induction times are used, which include a growth time chen2015 C5. In the felodipine, bifonazole and nimesulide growth studies nuclei are only formed and then grown: films are stored over P2O5 at 40 °C (or at ambient temperature) before the hot-stage growth runs, and nucleation with the polymers is not quantified kestur2010a P3, X10; kestur2011 P2. The spin-coating screen of vaneerdenbrugh2010b scores the crystalline area over 7 days, which mixes nucleation and growth vaneerdenbrugh2010b P2, C29. In the crystallization-from-solution papers by unseeded induction times (see nucleation induction time) and by the nucleation-rate ratio from seeded and unseeded runs [alonzo2012 P5, P10; ilevbare2013c P9]. In the physical-stability papers, by counting birefringent sites in films at fixed RH and temperature, and by the Arrhenius slope of the rate above and below Tg konno2008 P4; marsac2006a P5. A lag before detectable crystals in stored powders was read as delayed nucleation [rumondor2009b R17, C29]. In the ASD dissolution papers it is not measured directly but inferred from concentration curves, induction times, particle appearance and Raman lag times [konno2008a X4; ilevbare2013d R16; raina2014b C32].

Values in the library
materialvalue or classconditionspaper ids
nilutamide glass≈12.6 nuclei per sample at −30 °C; ≈0.8–2.5 at −20 to 40 °C30-min hold, open surface, 1 min growth at 65 °Ctrasi2014b R9
acetaminophennucleation zone 50–60 °CDSC cool–reheattrasi2012a C22
acetaminophen with polymersHPMCAS reduces nuclei to about a third; PAA gives nearly three times more10% w/wtrasi2012a C23
flutamidenucleation zone 10–40 °CDSC cool–reheattrasi2012b R10
nifedipine against felodipinefaster nucleation for nifedipinepure films, 22 °C, 0% RHmarsac2006a R1
felodipine filmsrate rises with storage RH; drug alone fastest22 °C, with polymerskonno2008 C11
indomethacin polymorphsδ fastest at 60 °C; α favoured at 80 °C; γ not readily (only from seeds); polymers retarded nucleationamorphous films, storage at 60 and 80 °Ctian2017 R1, R5, C7
celecoxibsamples "quite hard to nucleate"hot stagemosqueragiraldo2014 R15
vemurafenib βnucleation the limiting step above 150 °C (qualitative)hot stagelu2016 C46
pure nifedipine, felodipine against dispersionsnucleation much faster for the pure drugscited from a companion studymarsac2006b R16
indomethacin–PVPnucleus requires breaking the PVP–drug bond (proposed, untested)no crystallization datataylor1997 C43, X8
flufenamic acid–PSSA, chlorpropamide–PSSA, lidocaine–PVP filmsfully crystalline within 30 min of coatingspin coatingvaneerdenbrugh2010b R2.an, R2.s, R2.as
celecoxib; efavirenz; ritonavirinduction time about 2.4; 5.1; 120 minat the calculated amorphous solubilityilevbare2013c R8
felodipine, HPMC 1 µg/mLJ/J_0 7.05 × 10⁻⁴S 10[alonzo2012 R17]
danazol6–10 min without polymer; 88–516 min with 10 µg/mL polymer25 °C[jackson2014 R17]
nine drugs with carboxylated cellulose derivativescontact with the emerging nucleus "clearly a prerequisite" for disruption5 µg/mL, pH 6.8[mosqueragiraldo2018a C44]
amorphous felodipinerate ≈1.4 × 10⁵ → 2.9 × 10⁶ m⁻³ s⁻¹0 → 75% RH, 22 °Ckonno2008 R1.a, R3.a
nifedipine vs felodipinefaster at every PVP level and RH0–75% RHmarsac2008a R1, C7
felodipine with 50% HPMCASnegligible crystals for 35–120 days at 52–93% RH, read as delayed nucleationPXRD of powders[rumondor2009b R17, C29]
flutamide with 10 mol% PVAc, PIPAgrowth rate at the DSC exotherm maximum 7.7 and 7.0 against 4.4 µm/s, read as fewer nucleiDSC and growth curvetrasi2012b R11.b, R11.d, R11.f, C18
flutamide with 10 mol% nilutamideone nucleus in the imaged regioncooled to −30 °C, reheatedtrasi2012b R12, C16
felodipine with 3% PVP≈150-fold lower rate (cited)30 °Ckestur2013b R22, C35
ritonavir 17 against 20 µg/mL (below and above LLPS)induction about 284 against 119.3 minpH 6.8, 37 °C[ilevbare2013d R16, R12]
felodipine methanol shift with HPMC, S about 60precipitation not completely suppressed25 °C[alonzo2011 C30]
nifedipine α in γ slurries with polymerlag lengthened at similar slopes, read as α nucleation inhibition25 °Craina2014b C32
Pitfalls and disagreements

Nucleation is stochastic and often heterogeneous, which scatters crystallization temperatures baird2010 C19; baird2013 C18. Nuclei counts are not rates unless area and time are controlled trasi2014b X7. Polymers can act oppositely on nucleation and growth: HPMCAS was the best nucleation inhibitor but the weakest growth inhibitor for acetaminophen trasi2012a C46, and drug–additive hydrogen bonding correlated with growth inhibition but not with nucleation for flutamide trasi2012b C45. Classical nucleation theory does not capture molecular recognition or polymer effects trasi2012a C49. The role of the hydrogen bond in nucleation is argued, not tested taylor1997 X8. The growth-rate ranking of polymers says nothing about nucleation, which was not measured kestur2010a X10; kestur2011 P3. The film screen cannot separate crystallization during coating from crystallization during storage except through the day-0 score vaneerdenbrugh2010b R2. In solution, how additives slow nucleation is "more speculative" than how they slow growth, because the molecular events cannot be observed ilevbare2013c C9; the papers do not separate nucleation from early growth inside the induction time ilevbare2013c X14. The delayed-nucleation reading of the HPMCAS lag in stored felodipine powders assumes that growth below and above the detection limit follows the same factors [rumondor2009b C28]. Nucleation counts separated polymers at 33% RH that PXRD crystallinity could not separate within 490 days (synthesis) [konno2008 R2.b; rumondor2009b R19]. The flutamide nucleation verdicts rest on two Tmax values per sample and an unnamed test trasi2012b X4. Nuclei per cycle are not rates when the zones differ in width trasi2012a X8. The felodipine "nucleation then growth" reading rests on curves alone [konno2008a X4]; the ritonavir 17 µg/mL comparison has no n or SD and mixes droplets with a higher bulk supersaturation [ilevbare2013d X18].

← Concepts and formulas

concept · as used in Crystallization tendency

physical stability

The ability of an amorphous drug or dispersion to stay amorphous, and unseparated, during manufacture and storage vaneerdenbrugh2010a C19; kestur2013b C46. In this chapter it is approached through the crystallization tendency of the neat drug; dispersion stability is treated in other chapters. New aspects (Miscibility and drug-polymer interactions): For polymer dispersions, physical stability is read as the time to the first crystalline peaks under a stress condition, and it depends on whether the polymer bonds the drug. Naringenin and quercetin dispersions with E100 or PVP showed no crystals for more than 200 to more than 450 days at each of five storage conditions, whereas dispersions with HPMCAS, HPMC, CMCAB or PAA crystallized within 1–3 days at 84% RH/25 °C and at 75% RH/40 °C wegiel2013a R9, C22. Lumefantrine–PVPVA dispersions crystallized after 1 to 8 weeks at 40 °C/75% RH, while enteric-polymer dispersions showed drug crystals only at 50% drug loading hiew2022a R6, C10. Phase separation is expected to lower stability: a phase-separated dispersion "would be expected" to crystallize faster in storage and dissolution [purohit2015a C2, C35; li2016c C1, C41]. Dry storage, humidity and polymer content each change the ranking of polymers [wegiel2013a R9, X3; vaneerdenbrugh2011a C50]. New aspect (Physical stability): for the resveratrol and curcumin dispersions it is likewise read as days before crystals appear at a set storage condition, and it depended "greatly" on polymer and condition wegiel2013b C23; wegiel2014 C21. New aspect (Supersaturation from ASD dissolution): two settings in which physical stability decides dissolution performance. For dispersions with two polymers: at 60% indomethacin and 40 °C/75% RH the HPMC and the EPO binaries both crystallized within 5 days (growth much slower with EPO), whereas the EPO–HPMC ternary stayed amorphous for more than 100 days, contrary to the expectation that partly replacing EPO with HPMC would make it similar or slightly worse ("unexpected"); at 30% drug the ternary was also more stable than the HPMC binary [xie2017 R26, C51, R27]. For suspensions of dispersions dosed preclinically, the crystallization tendency "is expected" to depend on the extent of supersaturation, the polymer solubility in the medium and the polymer's ability to inhibit crystallization in solution and matrix [li2018 C60]. Lopinavir–HPMCP dispersions, with the strongest solubility suppression, were the most stable against crystallization ("correlates"), and where suppression brought the concentration to or below the crystalline solubility the authors see no driving force for crystallization, so that such dispersions "should remain" physically stable, dry and as suspensions [li2018 C61, C62, C63, C64].

Findings

Crystallization tendency: F17, F20, F21. Miscibility and drug-polymer interactions: F15, F17, F23, F24, F26. Physical stability: F24, F25, F26, F30, F33, F34, F35. Supersaturation from ASD dissolution: F18, F20, F23.

Related concepts

includes glass stability and surface crystallization; depends on crystallization tendency; related to amorphous salt; depends on drug-polymer interactions, acid-base interaction and amorphous-amorphous phase separation; includes crystallization inhibition; tested by accelerated stability testing; lost through moisture-induced crystallization; depends on solubility suppression in suspensions; related to ternary amorphous solid dispersion and matrix crystallization.

Formulas

none.

How it is measured or obtained

By storage at set temperature and humidity followed by PLM, XRPD or fluorescence baird2010 P7; tian2016 R17; hiew2022b R7; by Raman or PXRD crystallinity against time kestur2013b R3. For the lumefantrine and polyphenol dispersions: open-dish storage at 40 °C/75% RH with PXRD at times from 1 to 76 weeks hiew2022a P8; five conditions (22–25 °C over P2O5, 22–25 °C at 84% RH over potassium chloride, 40 °C at 75% RH over sodium chloride, 50 °C and 90 °C over Drierite) with PXRD for the first sharp peaks wegiel2013a P3; for thin films, polarized-light scoring of the crystalline area over 7 days of dry storage, expressed as an amorphicity index [vaneerdenbrugh2011a P2, P3, E1]. For resveratrol and curcumin, storage tables of first sampling times with crystals by XRPD at five or six temperature–humidity conditions wegiel2013b P3, P4, R12; wegiel2014 P5, R13. In the ASD dissolution papers: open-vial storage of 60% and 30% dispersions at 40 °C/75% RH with PXRD up to 100 days [xie2017 P16, R26]; in suspension, concentration over 48 h and polarized light microscopy of 50% dispersions in pH 3 buffer without a dissolved inhibitor [li2018 P6, P7].

Values in the library
materialvalue or classconditionspaper ids
indoprofen; droperidol, nifedipine; clotrimazole, felodipinecrystalline in 1 day; by 3 months; edges only at 3 monthsquenched glasses, 0% RH, 14–20 °C below Tgbaird2010 R23, R24, R25
felodipine with 3% PVPnot completely crystallized after 24 weeks30 °C, drykestur2013b C11
indomethacin–PVP dispersions, 10–90% drugamorphous over 3 months (all compositions)40 °C, desiccanttian2016 R16, C35
indomethacin with HPMC or HPMCAS, 10–90% drugabove the miscibility limit crystalline in 2–15 days; at or below it amorphous 13–14 months40 °C, desiccanttian2016 R17, C53
pretomanid–HPMCAS dispersionsstable several months; neat glass crystallized within minutesambientnguyen2023a C62
lumefantrine HCl and sulfate saltsno recrystallization for at least 16 weeks40 °C/75% RHhiew2022b R7
delamanid saltsno PXRD peaks after 2 weeks40 °C/75% RHduong2022a C32
naringenin, quercetin with E100 and PVPno crystals for >200 to >450 daysfive conditions, 40–50% polymerwegiel2013a R9
naringenin, quercetin with HPMCAS, HPMC, CMCAB, PAAfirst crystals after 1–3 days84% RH/25 °C and 75% RH/40 °C, 30–80% polymerwegiel2013a R9
lumefantrine–PVPVA at 50, 25, 10 and 5% drug loadingfirst crystals after 1, 2, 4 and 8 weeks40 °C/75% RH, open dishhiew2022a R6
lumefantrine at 50% drug loading with HPMCAS, Eudragit L 100, HPMCP, CAPfirst drug crystals after 1, 8, 32 and 40 weekssamehiew2022a R6
resveratrol with PVP K29/32, E100, HPMC, HPMCAS, CMCAB, PAAPVP never crystallized (> 400 days); E100 > 400 days at 84% RH; the others crystalline at day 1 at 84% RH25 °Cwegiel2013b R12
curcumin with E100, cellulosics, PVP; neat> 350, 75, 22 and 14 days at 84% RH25 °Cwegiel2014 R13
curcumin dispersionsall > 350 days at 0% RH/25 °C; all crystalline within 1 day at 90 °C/0% RH and 60 °C/98% RHstoragewegiel2014 R13, C23
indomethacin 60% with HPMC; EPO; EPO–HPMC 6:2.9:1.1crystals within 5 days; within 5 days, slower growth; amorphous beyond 100 days40 °C/75% RH, open vials[xie2017 R26]
indomethacin 30%, EPO–HPMC against HPMCternary more stable (text only)40 °C/75% RH[xie2017 R27]
lopinavir 50% HPMCP dispersion in suspension≈2.5–3.0 µg/mL over 48 h; no crystals at 48 hpH 3, 25 °C, no inhibitor[li2018 R6, R7]
lopinavir 50% EC; CA dispersions in suspension≈4.3–5.0 µg/mL to 12 h, then ≈1.6–3.4; ≈10.5–11.2 µg/mL to 12 h, then ≈2.3–3.9same[li2018 R6]
delamanid sulfonate salts, neatseven of eight amorphous with Tg 83.7–117.8 °C against 43.3 °C for the free base; all amorphous after 2 weeks at 40 °C/75% RH, four liquefiedopen dishduong2022a R20
delamanid salt dispersions at 20% drug with HPMCASTg 100.4–117.0 °C against 86.7 °C; water uptake ≈10–13.6% against ≈8.2% at 95% RH37 °Cduong2022a R23, R22
praziquantel dispersions at 20% drugHPMC E5 LV amorphous at 23 weeks; HPMCAS-MF small crystalline peaks by 21 weeks40 °C/75% RH[polyzois2024 R7]
Pitfalls and disagreements

Miscible dispersions were stable while phase-separated ones crystallized quickly tian2016 C53. Extrapolation of high-temperature growth data can overestimate stability kestur2013a C7. The class of the neat drug is only a first flag (see F17). The storage time reached by dispersions without crystals is not stated for lumefantrine, so the ranking is relative and no shelf life follows hiew2022a X9. Some dispersions crystallize only partly and then stop, which the authors do not fully understand wegiel2013a C23, C43. Peaks in stored CAP dispersions came from crystalline material in the polymer itself, not from the drug hiew2022a C11, R7. Small unexplained peaks appear in low-loading HPMCP traces after 64–76 weeks hiew2022a X10. The polyphenol dispersions differ in polymer content (30–80%) and in storage protocol from the lumefantrine ones, so the two sets are compared only in direction wegiel2013a X3; hiew2022a X9. In the resveratrol and curcumin tables, day values are first sampling times with unstated intervals and a 5–10% detection limit wegiel2014 X10; wegiel2013b X16. The mildest and the harshest conditions did not separate any polymer wegiel2014 C23. Long-term stability of the EC and HPMCP dispersions at 15 and 33% drug is inferred from 48-h concentrations; suspension and matrix stability were tested only at 50% drug, where the EC dispersion lost supersaturation after 12 h [li2018 X8]. The 30% indomethacin comparison is stated in the text only [xie2017 R27]. The praziquantel statement that both dispersions stayed stable for several months is not borne out by the peaks in the HPMCAS-MF sample [polyzois2024 X1].

← Concepts and formulas

concept · as used in Crystallization tendency

plasticization by water

The lowering of the glass transition temperature of an amorphous drug by absorbed water, taken in these papers as about 10 °C per 1% water ilevbare2013a C23, E15; indulkar2015 C39. A water-saturated drug-rich phase therefore has a wet Tg far below that of the dry amorphous drug, and its state at the test temperature (supercooled liquid or glass) follows from it (F2). New aspect (ASD process): in the coating study of four drug–polymer systems, water sorbed at the storage humidity lowers the glass transition of the powder, and the wet Tg onset minus the storage temperature (Tg − T, negative when stored above Tg) is the number compared with crystallization onset moseson2022a P6, E2, R12. Nucleation and growth speed up with humidity because sorbed water raises mobility and lowers the wet Tg, so a coating that works on the surface cannot rescue a powder whose bulk is mobile moseson2022a C41, C25. In dissolution, hydration lowers the Tg of a drug-rich bicalutamide layer, and the authors list mobility above the wet Tg among the reasons why not every ASD is susceptible to matrix crystallization moseson2022b C63. For hydrating posaconazole–HPMCAS the gastric stage lets crystals grow inside the wetted matrix after water uptake and plasticization (the authors' reading) [shah2024 C45]. New aspect (Physical stability): in drug–polymer dispersions water erases the Tg gain of a high-Tg polymer: at 75–76% RH felodipine and nifedipine dispersions with up to 25% PVP fell below the Tg of the dry drug konno2008 C16; marsac2008a C17. Plasticization is "directly related to the hygroscopicity of polymer", yet per gram of water it lowers the Tg of the pure drug most konno2008 C20; marsac2008a C19. New aspect (Supersaturation from ASD dissolution): the ASD dissolution papers use it in two places: to explain why wetted amorphous felodipine crystallizes faster at 37 than at 25 °C (dry Tg about 45 °C, just below 35 °C at 75% RH, cited) alonzo2010 M1, C34, and to read the low Tg of an isolated ritonavir drug-rich phase as a ritonavir-rich phase holding a few percent of water [ilevbare2013d C8]. New aspect (Release mechanism and LoC): the wet Tg of the drug-rich phase decides whether nanodroplets are glassy or liquid at the test temperature. Indulkar and co-workers (2017) estimate that water-saturated amorphous nifedipine droplets have a Tg near 17 °C (dry 42 °C, 2.5% water), which makes drug exchange between droplets and solution slow at 5 °C, the premise of their isotope test [indulkar2017b C16, M1, C13]. Ritonavir, with a wet Tg around 0 °C, is a supercooled liquid under release conditions [que2019 R28, C49; indulkar2019a M1]. For indomethacin the relevant wet Tg is "plausibly" that of the drug-enriched surface (a 70% dispersion with about 6% water, onset 18 °C) rather than that of the neat drug (estimated 29 °C) saboo2020 C52, R30, M1; by the same 10 °C per 1% rule every lumefantrine dispersion's wet Tg would lie below 37 °C hiew2022b C44.

Findings

Amorphous solubility and LLPS: F2. ASD process: F44, F38, F41. Physical stability: F5, F6, F9, F12, F33, F34. Supersaturation from ASD dissolution: F1, F9. Release mechanism and LoC: F5, F9, F18.

Related concepts

lowers the glass transition temperature; measured with water sorption; decides glass-liquid phase separation against LLPS; related to supercooled liquid. Raises surface molecular mobility; lowers the Tg that decides surface crystallization; sets how long an atomic layer coating protects a powder. Special case of plasticization; set by hygroscopicity; raises molecular mobility; related to matrix crystallization, drug-rich phase and moisture-induced crystallization. Gives the wet glass transition temperature.

Formulas

water plasticization rule of thumb, Tg − T, Gordon-Taylor equation.

How it is measured or obtained

DSC of the isolated drug-rich phase or of amorphous drug stored at high humidity, with water from gravimetric sorption or from drying the pellet ilevbare2013a P8, P9, P15; mosqueragiraldo2015 P4, P5; almeidaesousa2015 P8. For stored ASD powders the wet Tg onset was read in hermetically sealed DSC pans after 24 h at the storage humidity, 5 °C/min, triplicate preparations moseson2022a P6. The Tg of hydrated bicalutamide could not be determined [moseson2021 R27, X8]. In the felodipine and nifedipine moisture papers, DSC of bulk samples equilibrated at each RH and sealed konno2008 P6; marsac2008a P7, reported as ΔTg dry minus 75% RH konno2008 R9. In the ASD dissolution papers: by DSC of the phase isolated by ultracentrifugation [ilevbare2013d P9]; for felodipine the humid Tg is cited, not measured in the medium alonzo2010 C34. In the release-mechanism papers: dry Tg by DSC, water content extrapolated from a 25 °C sorption profile to 100% RH, and a rule of about 10 °C Tg depression per 1% water [indulkar2017b P3, P4, E6]. DSC of a 70% dispersion after 4 h at 100% RH saboo2020 P23; sorption at 37 °C extrapolated to 100% RH with the rule [yang2021 P9].

Values in the library
materialvalue or classconditionspaper ids
ritonavirTg 50.4 → 0.4 °C with 4.03–5.15% waterisolated phase, 10–50 °Cilevbare2013a R5, R6
clotrimazole; atazanavirTg 28 → about 14 °C (1.4% water); 104 → about 51 °C (5.2%)precipitates, 37 °Cindulkar2015 R22, M1, M2
telaprevirTg 103 → 52 °C; 4.31% water at 95% RHpellet, 37 °Cmosqueragiraldo2015 R2, M1
sorafenib; griseofulvinTg 88.2 → 49 °C; 90.8 → 88 °C at 97% RHamorphous drugalmeidaesousa2015 M5, M6, R9
erlotinib–HPMCAS 60% DLwet Tg − T about −8.8 °C (75% RH) and −2.3 ± 1.1 °C (43% RH)24 h at RH, T = 40 °Cmoseson2022a R12
erlotinib–PVPVA 50% DL−0.5 ± 1.2 °C43% RHmoseson2022a R12
naproxen–HPMCAS 10% DL; lumefantrine–PVPVA 40% DLabout +11.3 °C (75% RH); +10.7, +14.3, +11.3 and +11.4 °C (43% and 31% RH)samemoseson2022a R12
rule of thumb for stabilitymore than 50 °C between storage temperature and Tg (cited); the authors state the benefit of the coating holds at about 0 to +15 °Cmoseson2022a E3, C45, R17
felodipine with 25% PVP, HPMCAS, HPMCTg 54.9 → 31.1, 44.2 → 35.6, 43.1 → 34.4 °C0 → 75% RHkonno2008 R5
felodipine dispersions ≤ 25% polymerTg ≈30–35 °C after 75% RH, below dry felodipine ≈43 °C25 °Ckonno2008 R7, C16
nifedipine pure; 25% PVP≈43 → 26 °C; ≈54 → 29 °C0 → 76% RHmarsac2008a R6
amorphous felodipine (cited)Tg ≈45.7 → 29.6 °C, smooth0 → 94% RHrumondor2010 R10
pimozide–PVP 50:50wet Tgs −21 and 44 °C94% RH[rumondor2009c R17]
ketoprofen–PVP 70:30wet Tg −23 °C94% RH[rumondor2009c R25]
celecoxib 6:4 dispersionsTg 64–103 → 22–42.1 °C after 3 days at 94% RHDSC[xie2016b R17, R20]
tacrolimus; tacrolimus–HPMC 1:163.6 → 51.4 °C; 77.2 → 49.0 °C after 3 days at 75% RHDSCtrasi2017 R15
lumefantrine dispersions precipitated into water4–6% water (HPMCP, HPMCAS), 9% (CAP); Tg 45–112 °CTGA; DSC[bhujbal2021 R10]
felodipine (cited)Tg about 45 °C dry, just below 35 °C after 75% RHstorage dataalonzo2010 M1
ritonavir drug-rich phase from a PVP dispersionTg inflection 1.05 °C against 50 °C for the amorphous drugwater, 37 °C, DSC at 20 °C/min[ilevbare2013d R2, M1]
indomethacin (cited)Tg falls rapidly with water content, as for felodipineliteraturealonzo2010 R24
nifedipinedry Tg 42 °C; wet Tg approximately 17 °C (estimated)2.5% water (estimated)[indulkar2017b M1, C16]
ritonavirTg 50 °C; wet Tg around 0 °C (cited)release conditions[que2019 R28]
indomethacin; indomethacin–PVPVA 70%wet Tg 29 °C (estimated); onset 62.64 → 18.48 °C with ≈6% water37 °C, 100% RHsaboo2020 M1, R30
loratadine, miconazole, danazol, elbasvirwet Tg 13.8 (DSC), −4.7, 73.3, 129.1 °C (rule)37 °C[yang2021 M1, M2, M3, M4]
atazanavirdry 105 °C, wet 55 °Ccited[correasoto2022b M1]
Pitfalls and disagreements

The griseofulvin Tg falls only 2.8 °C with an estimated 2.7% water, against 39 °C for sorafenib with 1.8%, and storage time and water content are not stated almeidaesousa2015 X11. The water content of the telaprevir pellet was not measured mosqueragiraldo2015 X3. For the coated powders the wet Tg values are not printed, only Tg − T bars, and the definition in words is the reverse of the sign used moseson2022a R12, X24, E2. Uncoated powders failed in about 1–2 weeks across the whole range of Tg − T, so Tg − T orders the coated failures only in part moseson2022a X15. The 0 to +15 °C statement applies to coated powders only moseson2022a C45, R11. The rule of 10 °C per 1% water given in the Definition fits the pure drugs but not the 25% polymer dispersions of the felodipine study, which drop about 5 °C per 1% (synthesis: 23.7 °C with 4.68% water konno2008 R6). Plasticization did not by itself cause phase separation or rank nucleation [rumondor2009c C38; konno2008 C26]. Plasticization by water did not rank celecoxib dispersions at 94% RH: four Tgs within about 5 °C, very different stability [xie2016b C46]. The water content of the ritonavir phase was not measured [ilevbare2013d C8]. Plasticization does not rank crystallization: indomethacin, with a similar Tg drop, crystallized much more slowly than felodipine during dissolution alonzo2010 C42. The 17 °C nifedipine value is an estimate from a rule taken from indomethacin; no Tg of water-saturated nifedipine was measured [indulkar2017b X8]. The 70% indomethacin dispersion is a model of the surface layer saboo2020 X16, and most wet Tgs are rule estimates [yang2021 P9].

← Concepts and formulas

concept · as used in Crystallization tendency

plasticization

Lowering of the glass transition temperature of a mixture by a component with a lower Tg; in the library water is the usual plasticizer of drugs, polymers and dispersions, and drugs, surfactants and plasticizers lower the Tg of polymers [konno2008 C2, C14; marsac2008a C1; correasoto2022b C39]. The opposite, a higher-Tg component raising the Tg, is antiplasticization konno2008 C16. New aspect (Supersaturation from ASD dissolution): plasticization by water acts during dissolution as well as during storage. For delamanid salts, four of eight liquefied at 40 °C/75% RH, which the authors attribute "presumably" to plasticization by water, and the salts took up 5.8–21.1% water against far less for the free base duong2022a R20, C27. In the undissolved matrix of an enteric dispersion in acid, absorbed water lowers the Tg and raises molecular mobility, which the authors give as the reason why the drug can crystallize there although the medium is subsaturated; water content and Tg were not measured in that state [nguyen2023c C43]. In hot-melt extrusion the drug itself plasticizes the polymer, which the authors use to justify extruding a drug at the polymer's Tg ("would") [alvarenga2025 C46]. New aspect (Release mechanism and LoC): in the release papers of 2021–2022 plasticization of the dispersion by the drug or by an added surfactant is used to explain faster polymer release. Clopidogrel (Tg −9 °C) lowered the dispersion Tg, and the faster PVPVA release from its binary dispersions is called a plasticization effect, as for PMMA with 20 wt% PEO, whose dissolution rate rose by a factor of two (cited) [correasoto2022a C48, R30]. TPGS lowered the Tg of 30% felodipine–PVPVA dispersions and "could" raise the limit of congruency through higher molecular mobility [saboo2021 R5, C43]. Surfactants and VP lowered the Tg of 10% atazanavir–PVPVA dispersions, and improved release is "due, at least in part," to plasticization raising polymer-chain mobility [correasoto2022b R20, C40]. In the other direction, more low-polarity drug "would be anticipated" to reduce water penetration and water plasticization and so lower the polymer rate [yang2021 C32], and a drug–polymer interaction that persists in water is proposed to do the same [que2021 C44]. Surfactants can also promote crystallization by plasticizing the matrix (cited), and Tween 80 and TPGS "could possibly" plasticize ritonavir droplets so that they coalesce indulkar2022 C40, C48; except for SDS every surfactant lowered the single Tg of 30:70 ritonavir–PVPVA (midpoints 70.90–77.51 against 82.24 °C) indulkar2022 R18. Built into PVPVA dispersions of high-Tg drugs, plasticizers lowered the Tg and raised release and the limit; the authors suggest increased polymer chain mobility, that is a faster disentanglement rate, as one mechanism [correasoto2023 C30, C39]. Faster initial water entry from drug plasticization, together with phase separation moving most water into the hydrophilic phase, is a "possible explanation" for polymer release rising faster than water penetration in MePHPH–PVPVA up to 20% [deac2023a C51, C50]. The 2023 surfactant study attributes barrier disruption by non-ionic surfactants to plasticization: Span 20 and Span 85 lowered the Tg of amorphous ritonavir from 46.8 °C to 7.7–27.8 °C and, partitioning into the drug-rich phase, are taken to soften the drug-rich layer so that flow removes it yang2023 R22, C11, C18, C35. The 2025 high-loading particle study warns that SDS, also a plasticizer, may predispose amorphous formulations to phase separation and recrystallization [hiew2025 C58].

Findings

Physical stability: F6, F9, F22, F28, F32. Supersaturation from ASD dissolution: F23, F24. Release mechanism and LoC: F15, F18, F21, F26, F31.

Related concepts

special case plasticization by water; contrasts with antiplasticization; measured by glass transition temperature; raises molecular mobility; tested by Tg-scaled crystal growth rate; lowers physical stability and matrix crystallization; limits in situ salt formation. Speeds polymer chain disentanglement.

Formulas

Gordon-Taylor equation gives the ideal-mixing Tg that plasticization lowers.

How it is measured or obtained

DSC of samples equilibrated at a given RH and sealed hermetically, onset or midpoint Tg konno2008 P6; marsac2008a P7; the drop in Tg between the dry state and storage at 75% RH, ΔTg konno2008 R9. Outside the subcategory, Tg of drug–polymer–surfactant or plasticizer blends by DSC [correasoto2023 R17, C30; nguyen2023a R13]. In the ASD dissolution papers: water sorption at rising relative humidity with DSC of the same materials for Tg, and open-dish storage at 40 °C/75% RH with visual inspection and PXRD duong2022a P12, P10, P13; in extrusion, the drug Tg and the polymer Tg set the processing window [alvarenga2025 M1, M2, M3, E2]. In the release-mechanism papers: DSC of dispersions with and without the additive [saboo2021 R5; correasoto2022b R13].

Values in the library
materialvalue or classconditionspaper ids
felodipine with 25% PVP, HPMCAS, HPMCTg 54.9 → 31.1, 44.2 → 35.6, 43.1 → 34.4 °C0 → 75% RH, 25 °Ckonno2008 R5
felodipine–PVP 0–100%ΔTg ≈15 °C (7% PVP) to ≈158 °C (pure PVP)dry minus 75% RHkonno2008 R9
25% polymer dispersionsTg drop ≈13.3 °C at ≈1.1% water (PVP) vs ≈6.2 °C (HPMCAS)25 °Ckonno2008 R6, C14
nifedipine, pure and 25% PVP≈43 → 31 °C with ≈1.15 g/100 g water; ≈54 → 43 °C with ≈2 g/100 g25 °Cmarsac2008a R7, C19
pimozide–PVP 50:50two Tgs −21 and 44 °C wet against one at 94 °C dry94% RH[rumondor2009c R17]
PTM–HPMCAS–TPGS (outside)Tg ≈79 → 63 °C with TPGS at 20% loadingDSCnguyen2023a R13, C22
atazanavir and ledipasvir ASDs with plasticizers (outside)lower Tg than the binary ASDDSC[correasoto2023 C30]
celecoxib with 10% TPGS or sucrose palmitateTg 55.34 → 44.81 and 44.46 °C; growth fasterDSCmosqueragiraldo2014 R5, C16
pretomanid–HPMCAS with TPGS (other chapter)Tg falls as TPGS risesDSCnguyen2023a R13, C22
ritonavir–PVPVA with Span 20 or Span 85Spans in the drug-rich phase, read as plasticizing it and speeding separation ("likely")O-PTIR, NMR, confocal[yang2022b C58, X13]
griseofulvin, DPB, celecoxib with 5% PHBTg −5 to −12 °C; growth up to ≈10-fold fasterDSC; melt growthsato2016 R7, R11, X6
delamanid sulfonate salts, neatwater 5.8–7.1% (aromatic), 8.1–11.6% (camsylate, edisylate), 16.2–21.1% (esylate, mesylate); four liquefied95% RH; 2 weeks at 40 °C/75% RHduong2022a R20
delamanid edisylate 1:1 with excess acid59% water uptake and Tg 53.9 °C against 11.6% and 117.8 °C for the 2:1 salt95% RHduong2022a R21
delamanid salt dispersions at 20% drug with HPMCASwater ≈10–13.6% against ≈8.2% for the free-base dispersion; Tg 100.4–117.0 against 86.7 °C95% RHduong2022a R22, R23
delamanid free-base dispersions, 0 to 20% drugTg 121.0 → 86.7 °C (HPMCAS) and 120.1 → 82.0 °C (HPMCP); water at 95% RH ≈10.8–11.6% → ≈7.9–8.2%DSC, sorptionduong2022a R9, R10
praziquantel, racemate and R-enantiomerTg about 37 and 34 °C, used to place the extrusion temperatures at 150 and 120 °CDSC[alvarenga2025 M2, M1]
felodipine–PVPVA 30% with 0, 2, 5, 10% TPGSTg 89.2, 86.0, 80.8, 71.1 °C (felodipine 44.7, PVPVA 105.4 °C)DSC, dry[saboo2021 R5]
atazanavir–PVPVA 10% with SDS, CTAB, TPGS, Tween 80, Span 20, Span 80, Span 85, VPTg lowered by 16, 9, 14, 14, 11, 13, 7, 16 °C from the binaryDSC, dry[correasoto2022b R20]
clopidogrel–PVPVA 40–60%PVPVA ≈4.1–6.7 against ≈2.6–2.7 mg/min/cm² for neat PVPVAcompacts, 37 °C[correasoto2022a R18, C48]
loratadine–HPMCAS 5%HPMCAS released faster than from neat HPMCAS, "feasible" plasticization by a low-Tg drugcompacts[yang2021 C31, R18]
ritonavir–PVPVA 30:70 with 5 parts Tween 80, TPGS, Span 20, Span 85, SDSTg midpoint 70.90, 72.84, 73.24, 77.51, 84.93 °C against 82.24 °CDSC, dryindulkar2022 R18
8:1 atazanavir and ledipasvir glasses with nine plasticizersTg lowered by 30.4–43.3 and 47.6–53.1 °Cmodulated DSC[correasoto2023 R17, R18]
ledipasvir–PVPVA 10% with 5 and 10 g TEACTg 99.1 → 85.8 °C; release 20 → 94% by 90 minDSC; compacts, pH 6.8[correasoto2023 R9]
amorphous ritonavir with Span 20 (80–20, 60–40)onset Tg 14.8, 7.7 °C (46.8 °C neat)DSCyang2023 R22
amorphous ritonavir with Span 85 (80–20, 60–40)27.8, 12.7 °CDSCyang2023 R22

Evidence table: plasticization.

Pitfalls and disagreements

Two framings coexist: water is the more efficient plasticizer of PVP dispersions at equal water content (compared with cellulose dispersions) and the more efficient plasticizer of the drug alone per gram of water (compared with PVP dispersions); both hold, for different comparisons konno2008 C14; marsac2008a C19. A rule of thumb in the library puts the Tg drop at about 10 °C per 1% water [yang2021 E3], which fits pure drugs but overestimates the drop of the 25% polymer dispersions, about 5 °C per 1% (synthesis: 23.7 °C with 4.68% water for 25% PVP konno2008 R6). Plasticization alone does not cause moisture-induced phase separation [rumondor2009c C38]. Lower plasticization by water is offered as an alternative to the thermodynamic reading for HPMCAS dispersions [rumondor2010c C32, X12]. The Span plasticization rests on Tgs given only in the supplement [yang2022b X13]. Liquefaction was reported for four salts while the text and the table disagree on which, and the photograph of one looks unliquefied duong2022a X1, X11. The water content and Tg of the wetted matrix in acid, on which the mobility argument rests, were not measured [nguyen2023c C43]. For the extrusion window no Tg of a drug–polymer blend was measured [alvarenga2025 C46]. The dispersion Tg of the clopidogrel study is in the SI and not in the main text [correasoto2022a R26], and the loratadine–HPMCAS dispersion Tg was not measured [yang2021 C31]. Among the atazanavir additives the Tg depression and the release enhancement did not rank together (TPGS lowered the Tg by 14 °C and raised release only 4-fold, Span 80 by 13 °C and 35-fold) [correasoto2022b R20]. The Tg of the drug-rich layer formed in situ was not measured; plasticization is inferred from drug–Span mixtures yang2023 X17.

← Concepts and formulas

concept · as used in Crystallization tendency

polymer adsorption to drug-rich droplets

The accumulation of a dissolved polymer at the interface of the drug-rich droplets of a phase-separated solution, where it is thought to reduce droplet coalescence and to disturb heterogeneous nucleation at the droplet–water interface ("likely"); with the cellulosic polymers it is accompanied by some polymer mixing into the droplets [jackson2014 C33, C50, C53, C48]. The zeta potential of danazol droplets moved with the charge of the polymer over 1–1000 µg/mL (PVP −7.1 to −3.5, HPMC −2.9 to −2.2, HPMC-AS −8.9 to −18.6 mV against −8.1), and bile salts made telaprevir droplets about −40 to −53 mV against −9.2 mV [jackson2016 R18; lu2017 R8].

Findings

F15, F32.

Related concepts

part of liquid-liquid phase separation; related to droplet coalescence, zeta potential and polymer adsorption.

Formulas

none.

How it is measured or obtained

Indirectly: droplet size by DLS over hours, zeta potential of the droplets, and the lowered onset of phase separation; preliminary NMR gave the polymer content of the droplets (not shown) [jackson2014 R22, R23, R5, R26].

Values in the library
materialvalue or classconditionspaper ids
danazol droplets aloneabout 490 nm at once, about 9800 nm after 1 h25 °C[jackson2014 R22]
danazol droplets with HPMC; HPMC-ASabout 305–345 nm up to 2 h; about 230–250 nm up to 4 h10 µg/mL polymer[jackson2014 R22]
zeta potential: alone; PVP; HPMC; HPMC-AS−8.1; −5.3; −2.3; −13.7 mV25 °C[jackson2014 R23]
polymer in the dropletsaround 10–20% for HPMC, below about 10% for PVP, none for HPMC-ASpreliminary NMR[jackson2014 R26]
ritonavir nanodroplets (ASD dissolution chapter)charge repulsion between polymers adsorbed at the ritonavir–water interface "likely" stabilizes against coalescence37 °C[ilevbare2013d C38]
telaprevir droplets, 1.86 mM six bile saltszeta about −40 (SGC) to −53 mV (STCDC)180 µg/mL, pH 6.5lu2017 R8
Pitfalls and disagreements

The NMR is preliminary and not shown, with no value for HPMC-AS; the site of nucleation was not measured [jackson2014 X15]. The coalescence claim has no droplet size in the first minutes for danazol alone [jackson2014 X6]. Zeta potential was measured at 180 µg/mL, not in the induction-time solutions lu2017 X6.

← Concepts and formulas

concept · as used in Crystallization tendency

polymer adsorption

The attachment of a dissolved polymer to a surface, here the drug crystal (and, for danazol, the droplet interface). Every explanation of crystal growth inhibition in these papers assumes that the polymer must adsorb on the crystal to act, and "there is a general consensus" that it must (Mullin 2001, outside the library) [ilevbare2012c C3, C35; ilevbare2013b C47]. Adsorption can be physical (van der Waals, electrostatic, usually reversible) or chemical, and is driven by hydrophobic interaction, electrostatic attraction and hydrogen bonding [ilevbare2012c C4]. On crystal surfaces (formerly the page "polymer adsorption on crystal surfaces", from Crystallization from solution): the binding of a dissolved polymer to the faces of a growing drug crystal, which blocks growth sites and slows or shapes growth; it is usually face-selective, because molecular crystals are anisotropic, and changes habit when it slows the fast faces [ilevbare2012c C3, C5, R18]. It is the central assumed mechanism of crystal growth inhibition: the polymer occupies growth sites as a mechanical barrier and pins advancing steps [alonzo2012 C9; ilevbare2012c R19]. HPMCAS-LF was seen by AFM on three faces of felodipine, (1 1 −1), (1 −2 −2) and (1 −1 1), with no visible difference in density, taken as nonspecific adsorption, and AFM-IR identified the adsorbed material on a dried film [schram2015 R23, R25, R20, R21].

Findings

Crystallization from solution: F7, F8, F10, F11, F12, F23, F24, F27, F36, F38, F42, F44.

Related concepts

related to surface coverage, step pinning, polymer conformation, crystal habit and BFDH morphology prediction; mechanism of crystal growth inhibition.

Formulas

adsorbed amount by solution depletion, Langmuir-Freundlich isotherm, Kubota-Mullin model, Cabrera-Vermilyea step pinning.

How it is measured or obtained

By depletion: seeds equilibrated with polymer solution for 16–24 h, the amount adsorbed per area from the polymer lost from the supernatant, C_a = (C_i - C_f)\,V/(m \times A), with the polymer assayed by the phenol–sulfuric acid method at 490 nm, or by surface tension for HPMC [ilevbare2012b P9, E10; alonzo2012 P6]. In the ritonavir screen and the 2013 growth paper adsorption was not measured [ilevbare2012c C35; ilevbare2013b X14]. Adsorption on crystal faces is also read indirectly by growth-rate ratios against polymer concentration, by crystal habit and PXRD of the grown crystals, and by FTIR shifts of the drug bands after exposure to polymer [alonzo2012 R8; ilevbare2012c R4, R6; ilevbare2013b R12].

Values in the library
materialvalue or classconditionspaper ids
HPMC on felodipine seedsabout 60–167 µg/mL/m² up to 0.38 µg/mL equilibrium; 285–534 at 0.62–1.12 µg/mLsurface tension, 16–24 h[alonzo2012 R6]
CAP Adp 0.85 on ritonavir seedsplateau about 41 mg/m² (pH 3.8); about 27 mg/m² (pH 6.8)depletion, 4.0 mg seeds in 20 mL[ilevbare2012b R9, R10]
PVPVA on indomethacin crystalsglobule spacing 160 ± 50 nm (5 µg/mL); 70 ± 20 nm (50 µg/mL)AFM[moseson2020a R12]
HPMCAS on paclitaxel dendritesadsorption-site spacing about 10–16.8 nmSEM finger width[que2018 R12]
HPMCAS-LF pre-adsorbed on felodipine seedsovergrown by fresh drug in polymer-free solution (growth ratio about 1.8, then 1.08 in a second run)S 4, 25 °C[schram2016 R2, R3]
A5a, A5b, ECA, ECB, ECD on amorphous atazanavirA5a even, no globules; ECA, ECB small even globules; ECD larger; A5b large uneven globules5 µg/mL, 1 h, tapping-mode AFM in buffer[mosqueragiraldo2018a R4]
HPMCAS L, M, H on posaconazole dropletspolymer on every droplet surface from the ζ-potential ("is expected to delay surface crystallization")pH 6.5[shah2025 R15, C43, C44]
HPMC, felodipine seedsinhibition plateau (ratio about 0.55) where fitted coverage reaches about 1 (1 µg/mL)S 8, 25 °C[alonzo2012 R7, R8]
Pn-IPAAmd, ritonaviraspect ratio 4.4 against 8.8; (200) PXRD peak about 245 against 150 countsS 7.6, 37 °C[ilevbare2012c R4, R6]
PVPVA, celecoxib and efavirenz seedsN–H bands 3331 → 3321, 3225 → 3217; 3316 → 3308 cm⁻¹overnight in 100 µg/mLilevbare2013b R12
HPMCAS on paclitaxel seedspolymer slows but does not block growth at S 88seeded ASD dissolution[que2018 C30, C38]
HPMCAS, earlier felodipine work"may adsorb at the crystal surface and block growth sites"dilute supersaturated solutionrumondor2010 C38
felodipine, HPMCAS-LF, pH 3adsorption on three faces exposing methyl/oxygen, chlorine/methyl and methyl/amine/oxygen groupsAFM in liquid[schram2015 R23, R25]
HPMC-606, HPMCAS-MF on atazanavir with SDSstrong adsorption keeps SDS off the crystal ("we can speculate"), not measured1 mg/mL polymer, 0.1 mg/mL SDS[indulkar2019b C43]
bile salts on atazanavir needlesblocking of the needle sides "hypothesized" from aspect ratios 41–83 against 12.4biorelevant media[elkhabaz2021 C47, R21]
Pitfalls and disagreements

Adsorption by depletion has no crystal-free control, so loss by aggregation or on vessel walls would count as adsorption [ilevbare2012b X15]. The amount adsorbed did not predict the effect: more adsorbed at pH 3.8 and less inhibition [ilevbare2012b R9, R7]. Maximal adsorption did not stop growth at S 7.6 with 100 µg/mL polymer [ilevbare2012b R17]. Isotherm fits missed adsorption at low concentration [alonzo2012 C21]. The adsorbed amount was not measured in either felodipine–HPMCAS paper [schram2015 X7]. The AFM coverage ranking is qualitative, one image per polymer [mosqueragiraldo2018a X16]. The face-specific binding of Pn-IPAAmd rests on means of 50 crystals with overlapping spreads and PXRD relative intensities, with no adsorption or face-specific rate measured [ilevbare2012c X12]. The BFDH fast faces are assumed to be the fast faces of the real crystals [ilevbare2012c C45]. The PVPVA hydrogen bond is proposed from N–H shifts alone ilevbare2013b X13. Nonspecific adsorption is a visual judgement and the (1 −1 1) region shows streaks [schram2015 X8]. Face-selective adsorption is the stated general case for ritonavir [ilevbare2012c C5], whereas HPMCAS-LF on felodipine showed no visible difference in density between three faces [schram2015 R23, R25]; neither paper measured face-specific adsorbed amounts (synthesis: contrast between papers).

← Concepts and formulas

concept · as used in Crystallization tendency

polymer chain stiffness

How rigid a polymer backbone is

The authors attribute the advantage of cellulose derivatives over most synthetic polymers of similar hydrophobicity "most likely" to their semirigid, bulky backbones, which adsorb flat with more contact, against flexible chains that adsorb in loops with little contact [ilevbare2012c C39]. For nucleation the same property is given as a "perhaps" reason why bulky cellulose esters disrupt solute packing more than flexible chains ilevbare2013c C39.

Findings

F8, F13.

Related concepts

related to polymer conformation, conformational flexibility, polymer hydrophobicity and polymer adsorption.

Formulas

none.

How it is measured or obtained

Not measured. The argument rests on the classification of the polymer chemistry (cellulose against vinyl-type chains) and on a molecular-dynamics result from the literature that stiffer chains adsorb more easily [ilevbare2012c R16, C39].

Values in the library
materialvalue or classconditionspaper ids
synthetic polymers on ritonavir growthabout 0.9–1.4 except Pn-IPAAmd (3.25)5 µg/mL[ilevbare2012c R9]
cellulose-based polymers on ritonavir growthup to 2.985 µg/mL[ilevbare2012c R9]
PAlAmn (SP 20.70) and Pn-IPAAmd (22.64) on ritonavir and efavirenz induction timesabout 112 and 119 min (ritonavir), 4.4 and 5.0 min (efavirenz), against 172–244 and 21.6–28.0 min for adipate esters of similar SPinduction timeilevbare2013c R9, K4
Pitfalls and disagreements

Only two synthetic polymers fall in the matching SP range for the nucleation argument, and for celecoxib the synthetic Pn-IPAAmd was among the best ilevbare2013c K4, X11. Rigidity was not measured, and the SP values of the synthetic polymers are not printed in the growth paper [ilevbare2012c X11]. Pn-IPAAmd is flexible yet the best growth inhibitor, attributed to a specific interaction [ilevbare2012c C44, C47].

← Concepts and formulas

concept · as used in Crystallization tendency

polymorphism

The ability of a molecule to crystallize in more than one crystal structure towler2007 C2; the metastable form often crystallizes first, and recrystallization from the undercooled melt "frequently" gives a metastable polymorph baird2010 C16. This draft is written from the crystallization-tendency chapter; the evidence table lists further papers that later chapters will add.

Findings

F4, F7, F8.

Related concepts

includes monotropy; tested by Burger-Ramberger rules; related to cold crystallization, conformational flexibility and prenucleation clusters.

Formulas

Burger-Ramberger heat-of-fusion rule, Hoffman equation.

How it is measured or obtained

By Raman (including in liquid N2), XRPD, FTIR, DSC melting and solid-state NMR baird2010 P5; trasi2014b R15, R20, R25; lu2016 R18, R19; the polymorph produced is mapped against temperature or supersaturation lu2016 R10; towler2007 R1.

Values in the library
materialvalue or classconditionspaper ids
indomethacin α and γα above a supersaturation of 4, γ below about 3.5ethanol (cited)towler2007 R1
anthranilic acidform II from most solvents below 50 °C; form III from the melt or above 50 °Ccitedtowler2007 R2
vemurafenibat least four forms: α, β, γ, δsupercooled liquidlu2016 C41
nilutamidestable form, forms I and II, a suspected hydratemelt and humiditytrasi2014b R15, C25
indomethacin δ, α, γdifferent maximum growth rates and nucleation temperaturespure melttian2017 R7, R1
8 of 23 class I moleculesa different polymorph after liquid-N2 quenchRamanbaird2010 R3
Pitfalls and disagreements

The polymorph obtained from solution is sometimes stated without an identification method towler2007 X5. Forms γ and δ of vemurafenib rest on Raman and morphology only lu2016 X6. Solution motifs do not reliably foretell the polymorph towler2007 C33.

← Concepts and formulas

concept · as used in Crystallization tendency

prenucleation clusters

Aggregates of solute molecules in solution before a stable nucleus forms; clustering is taken as a prime requirement for crystallization, and if their structure could be tied to the final solid the polymorph "might" be controlled towler2007 C1, C2. Competing views hold them solvent-rich and disordered, polymorph-like, or a dense liquid that need not resemble the crystal towler2007 C4.

Findings

F8.

Related concepts

precedes nucleation; related to polymorphism and classical nucleation theory.

Formulas

none.

How it is measured or obtained

Inferred from concentration-dependent IR and Raman bands of the solution towler2007 P4, P5, R13; not measured directly.

Values in the library
materialvalue or classconditionspaper ids
indomethacincarboxylic acid dimer bands grow with concentration, even below saturationethanol, nitromethanetowler2007 R13, R15, R17
indomethacin in ethanol1660 to 1680 cm⁻¹ shift read as solute-rich clusters excluding solventIRtowler2007 C21
anthranilic acid in tolueneNH2 bands like form III, read as clusters unlike the crystallizing form IIIRtowler2007 C17, R9
Pitfalls and disagreements

Cluster size and composition were not measured; a constant band position fits single solvated molecules as well as clusters towler2007 X9; a bulk spectrum cannot exclude a minority population towler2007 X10.

← Concepts and formulas

concept · as used in Crystallization tendency

quenched-in nuclei

Nuclei formed while a melt is cooled faster than the rate that avoids detectable crystallization but slower than the rate that avoids nucleation; they cannot grow before vitrification and remain in the glass baird2010 C50; baird2013 C27.

Findings

F3, F16.

Related concepts

part of nucleation; defined through critical cooling rate; lowers glass stability; detected through critical heating rate.

Formulas

critical cooling rate to avoid nucleation.

How it is measured or obtained

Not observed directly in the library; inferred from crystallization on reheating and from the effect of the prior cooling rate on the critical heating rate baird2013 C28, C30, X7, or proposed from the class behaviour baird2010 C55; trasi2012a C40.

Values in the library
materialvalue or classconditionspaper ids
dibucaineRcrit,heat above 1000 °C/min after 20 °C/min cooling, 300 after 800 °C/minrapid heat–cool DSCbaird2013 R9.j, R11
clofoctolmelting endotherm after 20 °C/min cooling, none after 860 °C/min, at 200 °C/min reheatingrapid heat–cool DSCbaird2013 R12
class II and quench-glassy class I"most likely" explanation of recrystallization on reheatingDSC classesbaird2010 C55
acetaminophensuggested; nuclei from cooling not separated from those formed on reheatingDSC and hot stagetrasi2012a C40, X9
Pitfalls and disagreements

The nuclei were never seen or counted; the clofoctol test assumes equal nucleation during constant cooling baird2013 X7, C29. Their role in storage below Tg is proposed, not tested baird2010 C57; baird2013 X15.

← Concepts and formulas

concept · as used in Crystallization tendency

reduced glass transition temperature

T_{rg} = \frac{T_g}{T_m} with both temperatures in kelvin baird2010 E4. Assuming equal viscosity at Tg, a higher Trg means a more viscous liquid between Tg and Tm and so, it was expected, a better glass former; a typical value of 0.66 had been proposed baird2010 C1, R29.

Findings

F5, F12.

Related concepts

depends on glass transition temperature; related to Tm − Tg temperature gap; proposed predictor of glass-forming ability; contrasts with melt viscosity as a predictor.

Formulas

reduced glass transition temperature.

How it is measured or obtained

From DSC Tg (midpoint) and melting peak baird2010 P1; where Tg cannot be measured, Tg is estimated from a Trg–molecular-weight fit or a fixed Tg/Tm ratio baird2010 E27; vaneerdenbrugh2010a E7.

Values in the library
materialvalue or classconditionspaper ids
class I (23 molecules)0.69 ± 0.02DSCbaird2010 R14
class III (17 molecules)0.73 ± 0.04, not significantly different from class IDSCbaird2010 R14
classes II and III0.65–0.76 and 0.65–0.81DSCbaird2010 F5
ritonavir0.81DSCbaird2010 R18.ab
class II compounds0.65–0.76, not the constant 2/3DSCbaird2012 R14
pretomanid; delamanid0.67; 0.64, both class IDSCnguyen2023a C59
35 compoundsT_{rg} = 0.00021\,\mathrm{MW} + 0.64433,\quad R^2 = 0.44regressionbaird2010 E27, R19
Pitfalls and disagreements

Trg did not separate the classes, nor class I-A from I-B baird2010 C23, C37, and compounds with similar Tm − Tg had very different tendencies baird2012 C36. Trg of the 16 class I molecules without measured Tg is a function of molecular weight alone baird2010 X17. A small Tm − Tg (high Trg) did go with slow growth trasi2014a R8.h.

← Concepts and formulas

concept · as used in Crystallization tendency

secondary nucleation

The formation of new crystals in the presence of existing ones, from their surface or from fragments of them, with a lower energy barrier than primary nucleation [purohit2019 C3; duong2022b C35]. In the dissolution papers it enters in two ways: as the reason why seeds in an ASD speed crystallization and desupersaturation beyond their own growth [purohit2019 C3, C32], and as a process assumed to be small, so that the fall of the donor concentration reflects growth of the seeds alone [hate2019 C42]. New aspect (Crystallization from solution): it adds to the loss of dissolved drug in a seeded run and would confound a growth measurement [ilevbare2012a C14, C15; ilevbare2013b C12]; in seeded runs the authors judged it absent for celecoxib and ritonavir and present for efavirenz above 10 µg/mL ilevbare2013b R5, C13. Turbidity at 370 nm marks it in the telaprevir seeded runs (it rose at δ 7.9 and 12.5, not at 2.4), and STC and STCDC are said to inhibit it because no rise was seen (traces not shown); in acetaminophen PVPVA "does not appear" to stop it [lu2017a R4, R5, C22; trasi2015 C22].

Findings

Bioperformance: F29, F30. Crystallization from solution: F3, F5, F20, F27, F33, F42.

Related concepts

follows from crystal seeding; competes with crystal growth; part of desupersaturation; lowered by an absorptive sink; part of nucleation; a confounder of crystal growth rate; related to heterogeneous nucleation and residual crystallinity.

Formulas

power-law crystal growth rate (the fitted order assumes minimal secondary nucleation).

How it is measured or obtained

Not separated from growth in the Bioperformance papers. Elsewhere: turbidity in seeded runs, the size distribution of the seeds before and after growth, a mass balance of seed volume against solute removed, SEM of small crystals on larger ones, and the rise of a fine-particle count after a pH shift [ilevbare2012a R7, R8; ilevbare2013b R5, R6; que2018 R9, R10; shah2024 R14, C28]. In the seeded growth papers the turbidity is read at 280 or 350 nm, with an unseeded control at the same supersaturation and a volume balance on the seeds [ilevbare2013b P11, R5; ilevbare2012a R7, R8, R9].

Values in the library
materialvalue or classconditionspaper ids
tacrolimus seeded solutionsassumed minimal, so the desupersaturation rate gives the growth order (1.677)hollow-fibre module, S 8–20[hate2019 C42, R18]
ritonavir, celecoxib; efavirenzno significant secondary nucleation in seeded runs and no turbidity rise between 5 and 22 µg/mL (ritonavir also judged by size distribution and mass balance); efavirenz turbidity rose at every concentration above 10 µg/mL (S 1.2) once macroscopic crystals formedseeded, 37 °C; turbidity, size distributionilevbare2013b R5, R6, C12, C13
telaprevirsecondary nucleation with growth at δ 7.9 and 12.5, none at 2.4; inhibited by 1.86 mM STC or STCDC; extinction without bile salt 0 (δ 2.4), about 0.047 (δ 7.9), about 0.061 (δ 12.5)370 nm extinction, pH 6.5, 37 °C[lu2017a R4, R5, C11, C22]
paclitaxel seedssmall needles on larger crystals in SEM; lower aspect ratio after growthno polymer[que2018 R9, R10, C43]
posaconazole 10 wt% ASDfine-particle count rose after 15 min following the pH shiftFBRM and SHG[shah2024 R14, C28, C47]
ritonavir seeds after growthmonomodal size distribution; unseeded 10 µg/mL no loss in 8 hS 7.6[ilevbare2012a R7, R9]
telaprevir seeds, δ 7.9possible; bulk growth assumed dominant50 mM phosphate pH 6.5, 37 °C[lu2018 C8]
atazanavir needles from buffersecondary growth and small needles on larger ones; none in human fluidSEM, 300 min[elkhabaz2021 R17]
posaconazole with 0.4–20 µg/mL nanocrystal seedsinduction period removed; immediate desupersaturation at 2 and 20 µg/mLPBS pH 6.5, S 33.6duong2022b R13, C35
Pitfalls and disagreements

Secondary nucleation was assumed small in the tacrolimus growth fit and not tested [hate2019 C42]. The primary and secondary routes are not separated by induction times or particle counts trasi2015 X7. Breakage of seeds gives fine needles that look like secondary nuclei [que2018 C45]. In vivo, absorption lowers the supersaturation that drives secondary nucleation, and its role in the dog study is only argued [purohit2019 C41]. At concentrations near the amorphous solubility the 2-min slope of a seeded run may include secondary nucleation or a drug-rich phase, and no measurement separates them [ilevbare2012b X18]. The volume balance on the ritonavir seeds differs by a factor of 2.3 [ilevbare2012a X10]. For telaprevir secondary nucleation was inferred from turbidity, not measured on its own [lu2017a X10].

← Concepts and formulas

concept · as used in Crystallization tendency

solution crystallization

Nucleation and growth of crystals from the supersaturated solution, as against crystallization inside the undissolved ASD (matrix crystallization) [elkhabaz2017 C38; mosqueragiraldo2018b C52]. Crystallization from solution decided the enzalutamide-PVPVA dispersions (fall after the maximum) and the pretomanid case, in which droplets present above the amorphous solubility defeated HPMCAS-HF within minutes [wilson2018 C18, C42; nguyen2023a C13]. New aspect (Supersaturation from ASD dissolution): crystals nucleate and grow in the supersaturated liquid after the amorphous solid has dissolved, the second route by which an amorphous solid loses its advantage alonzo2010 C47. Polymer in a dispersion can move crystallization from the matrix to this route (nifedipine) raina2014b C21. When dispersions release completely, the authors of a celecoxib study take matrix crystallization as suppressed and attribute the differences between polymers to crystallization from solution, which the induction time measures xie2015 C39, C40.

Findings

Bioperformance: F9, F10, F11, F12, F14, F23, F27. Supersaturation from ASD dissolution: F1, F3, F4, F6, F14, F21, F24.

Related concepts

part of desupersaturation; contrasts with matrix crystallization; related to crystal growth and crystallization tendency; part of crystallization during dissolution; measured by nucleation induction time.

Formulas

supersaturation ratio, classical nucleation theory.

How it is measured or obtained

Fall of concentration with SHG, PLM or PXRD for the crystals; induction time by UV extinction [elkhabaz2017 P5, P8, P9, P10; hate2017 P2]. In the ASD dissolution papers: by a fall of the solution concentration together with crystals seen in the liquid rather than in the solid: needles between amorphous particles under polarized light alonzo2010 R19, fine plate-shaped crystals in the solution around dissolving dispersion particles raina2014b R13, free-floating needles in suspensions [chen2015a R15], or a rise of a pyrene ratio as crystals expel the probe [jackson2015 C14].

Values in the library
materialvalue or classconditionspaper ids
ezetimibe, buffer 21 µg/mLinduction about 1 min; PVP 100 µg/mL about 71 min; HPMC-AS about 50 min37 °C[elkhabaz2017 R6]
ezetimibe, FeSSIF 123 µg/mLHPMC-AS 50 µg/mL more than 360 min; PVP about 10 min37 °C[elkhabaz2017 R7]
nevirapine 500 µg/mLabout 5 min; PVPVA about 60 min; HPMC-AS over 4 hpH 6.5[hate2017 R2]
telaprevir 90% PVPVA ASDbelow 90 µg/mL from about 43 min; about 20 µg/mL at 100 min[mosqueragiraldo2018b R5]
atazanavir 50:50 PVPVA ASDcrystals from about 80 minhollow fibre[hate2018 R14]
indomethacin–Eudragit EPO ASDcrystallization after 15–20 min; small α crystals within about 20 minacid stage, pH 2.2, 37 °C[xie2017 R11, R23]
celecoxib, no polymer or PAAwithin 5 min at 22 µg/mL; ≈3.6 µg/mL at 1 hpH 6.8, 37 °C[xie2016a R2, R3]
enzalutamide 70 µg/mLcrystallized in about 15 min without polymer; held more than 16 h by four of 24 polymers added at 50 µg/mLpH 6.5[wilson2020 R1, C32]
pretomanid 80 µg/mL1.6–3.1 min with most enteric polymers, 22–24 min with HPMCAS-MF or HPMC, over 1000 min with HPMCAS-HF100 µg/mL polymer, pH 6.5nguyen2023a R5
amorphous indomethacinneedles in the liquid within 60 min, solid amorphous ≥4 h; ≈14.9 → ≈2.7 µg/mLpH 2, 25 °Calonzo2010 R19, R11, R7
nifedipine 50:50 dispersionscrystals below 2 µm in solution against 1–300 µm by the matrix routepH 6.8raina2014b R13, R2
danazol dispersions PVP / HPMC / HPMCASinduction ≈56–70 / ≈188–200 / ≈284–319 minpH 6.8, 25 °C[jackson2015 R5]
celecoxib–HPMCAS suspension with SDS; polysorbate 80free needles at 0.5 h; 1 hpH 3, 25 °C[chen2015a R14, R15]
celecoxib–PVP-K12 50% dispersiondesupersaturation about 2 h after complete releasepH 6.8, 37 °Cxie2015 C20
tacrolimus 50 µg/mLonset within 3 h; 6–8 h with HPMCpH 4.5, 37 °Ctrasi2016 R9
delamanid salt dispersions in acidconcentration did not generally fall during the acid stage, and 1 mg/mL predissolved PVPVA left release after a pH 3.0 stage unchangedpH 1.6–5.0, 37 °C[nguyen2023c R23, R22]
Pitfalls and disagreements

In the telaprevir and nevirapine solutions crystallization is inferred from concentration, without solid-state analysis [mosqueragiraldo2018b X13; hate2017 C42]. Crystallization of the enzalutamide-PVPVA solutions is inferred from concentration, without a solid-state analysis [wilson2018 X12]. For felodipine with dissolved polymers this route was named dominant without monitoring the solid in the vessel alonzo2010 X7. In the felodipine solvent-shift tests of 2008 no precipitate was examined [konno2008a X4]. The danazol crystallization is inferred from UV and pyrene, without solid-state identification [jackson2015 C14]. Complete release was taken to exclude matrix crystallization although the HPMC-containing celecoxib dispersions neither released fully nor were checked xie2015 X3. Where the medium is subsaturated, as in the gastric stage of an enteric dispersion, the solution route is excluded and crystals must have formed in the wetted solid [elkhabaz2019a C34; nguyen2023c C39].

← Concepts and formulas

concept · as used in Crystallization tendency

solvent evaporation rate

How fast the solvent is removed when an amorphous solid is made from solution; fast removal (spin coating) shortens the time the solution spends supersaturated before it becomes viscous, slow removal lengthens it vaneerdenbrugh2010a C11, C27.

Findings

F14.

Related concepts

affects crystallization tendency and its crystallization tendency classification after solvent evaporation.

Formulas

none (film thickness from weight gain in the chapter table).

How it is measured or obtained

Compared by method rather than measured: spin coating at 8000 rpm for 20 s against evaporation in an open vessel at ambient conditions vaneerdenbrugh2010a P2, P7; spin coating against rotary evaporation in later studies wegiel2013a R7; wegiel2014 C9.

Values in the library
materialvalue or classconditionspaper ids
31 slow-crystallizer combinations22 fully crystalline at day 0 after slow evaporationvial at ambient conditionsvaneerdenbrugh2010a R13
naringenin; quercetinamorphous naringenin by spin coating only; quercetin by neitherspin coating, rotary evaporationwegiel2013a R7
curcuminamorphous by spin coating, not in larger amounts by rotary evaporation—wegiel2014 C9
felodipine filmsthicker films from the more volatile DCM above 0.26 Mspin coatingvaneerdenbrugh2010a R1, C15
Pitfalls and disagreements

The drying time itself was not measured; the explanation that fast drying suppresses solvent effects is inferred vaneerdenbrugh2010a C25. Rotary evaporation also runs slightly warm (35 °C), which can promote crystallization vaneerdenbrugh2010b C39.

← Concepts and formulas

concept · as used in Crystallization tendency

spherulitic growth

Growth of a crystal as a round aggregate of radiating fibres (a spherulite) into the supercooled liquid; its diameter grows linearly with time and the slope is the growth rate kestur2010a C9, R4; kestur2011 R4. In the felodipine, bifonazole and nimesulide studies all crystals grew as spherulites, with and without polymer kestur2010a R2; kestur2011 R2. New aspect (Physical stability): felodipine crystals from amorphous thin films grow as spherulites below about 80% RH and switch to needle-like crystals of the same form at higher RH, with a 20–40-fold faster growth rumondor2010 R2, R8, C9, X2.

Findings

Miscibility and drug-polymer interactions: F10, F13. Physical stability: F3, F9, F19, F22, F27, F28, F29.

Related concepts

part of crystal growth rate and a form of its measurement; observed in supercooled liquid; related to polymorphism and moisture-induced crystallization.

Formulas

none.

How it is measured or obtained

By time-lapse polarized-light microscopy on a hot stage; the polymorph is checked by Raman microscopy and melting point kestur2010a P4, P5, R3; kestur2011 P4, R3. In the humid felodipine films, by photomicrographs of crystals at each RH and Raman of the two morphologies rumondor2010 P5, P6.

Values in the library
materialvalue or classconditionspaper ids
felodipine form II, neat and with 3% PVP, HPMCAS, PVP/VA, PVAcsmooth interface to the amorphous matrix in all five systems80 °Ckestur2010a R2
bifonazolerough fibre-like edges at 18 °C, smoother at higher temperature, sharper edges at 90–100 °Cno polymer; polymers did not change the profilekestur2011 R2
nimesulideneedle-like at 18–40 °C, zig-zag edges at 40–60 °C, smooth aboveas abovekestur2011 R2
felodipine, spin-coated films with polymers (other chapter)irregular boundaries and polymer-rich dark points on the crystallites22 °C, 3% HPMCAS, PVP, HPMCkonno2006 R2, C11
felodipine with PVP K-12, K-29/32, K-90 (other chapter)no significant change in macroscopic morphology0.5–4.5 wt%kestur2010b R5, R12, C8
amorphous felodipine filmround spherulite at 75% RH; needle-like crystal at 93% RH; same Raman peak positions25 °Crumondor2010 R2, R8
felodipine with HPMCASdisrupted spherulites with irregular boundaries, most at 10%32–93% RHrumondor2010 R4
felodipine with PVPseparate spherulitic segments at 64% RH and below; round outline from 75% RH2.5–10%rumondor2010 R6
felodipine form Icompact spherulites below 40 °C, rough edge at 45–65 °C, zigzag above 70 °C (surface)melt filmskestur2013a R5, R6
acetaminophen with seven polymersspherulitic shape kept10% w/wtrasi2012a R4
celecoxib with TPGS and PVPa large faceted plate at 120 °C instead of a fibrous frontmelt filmsmosqueragiraldo2014 R13, C34
ATP(R), ATP(S) with 5% polymersspherulites in all; size linear in timemelt films[sato2015 R3, R4]
flutamide, DPB with 5% PHBspherulites with a sharp interface, with and without polymermelt filmssato2016 R2
indomethacin δ and α (spherulites), γ (polycrystalline)morphology identifies the formmelt filmstian2017 R2

Evidence table: spherulitic growth.

Pitfalls and disagreements

The captions of the bifonazole–nimesulide morphology figure label both drugs' panels a to c, and the row assignment is inferred; the text describes bifonazole at 90 and 100 °C, which are not shown kestur2011 X3. The spherulite diameter slope is called a change in radius in one place of the felodipine paper kestur2010a X3. The linear size–time relation is assumed to hold for every system, but only single series are plotted for 80 °C kestur2010a R4. The Fig. 2 caption of the humid-film growth paper calls both crystals spherulitic although the 93% RH one is needle-like rumondor2010 X6. At about 84–86% RH both morphologies coexisted rumondor2010 X13. Because the felodipine front changes habit with temperature, rates at different temperatures do not refer to the same front kestur2013a X12.

← Concepts and formulas

concept · as used in Crystallization tendency

step pinning

The mechanism (Cabrera and Vermilyea, outside the library) by which adsorbed impurity molecules hold back an advancing growth step: the step must bulge between adsorbed molecules, its radius of curvature is limited by their spacing, closer spacing means slower growth, and growth stops when the spacing equals twice the critical radius \rho_c of a two-dimensional nucleus; because \rho_c falls as supersaturation rises, the maximum effect of a fixed impurity spacing falls with S [alonzo2012 C9, E8]. Ilevbare and co-workers chose it among four mechanisms for adsorbed impurities (surfactant action, incorporation, kink blocking, step pinning) by argument, because large polymers adsorb irreversibly [ilevbare2012b C12]. AFM of felodipine seeds shows steps held at several points by dissolved HPMCAS (spacing 68.5 ± 9.4 nm on buffer-grown seeds, 15.8 ± 7.4 nm on dispersion-grown seeds, step height 0.8 nm) [schram2016 R5, R12].

Findings

F9, F10, F23, F24, F12, F42.

Related concepts

part of polymer adsorption on crystal surfaces; depends on critical radius of a two-dimensional nucleus; related to impurity effectiveness factor, Kubota-Mullin model and kinetic roughening.

Formulas

Cabrera-Vermilyea step pinning, Kubota-Mullin model, Kubota-Mullin impurity effectiveness factor.

How it is measured or obtained

Not observed directly in these papers. It is tested through the fall of the growth-rate ratio with coverage, the factor α of the Kubota–Mullin model and the rise of the growth order from about 1.5 to about 2 [alonzo2012 R9, R13; ilevbare2012b R16]. In the ASD dissolution chapter, SEM finger width of paclitaxel dendrites with HPMCAS gives the spacing of adsorption sites (about 10–16.8 nm), and the distance between step pinnings on indomethacin crystals with PVPVA was 270 ± 100 nm [que2018 R12; moseson2020a R13].

Values in the library
materialvalue or classconditionspaper ids
felodipine with HPMCα 0.7, 0.5, 0.4, 0.6, 0.2; growth order 1.50 → 1.97 ± 0.12S 3–12, 25 °C[alonzo2012 R9, R13]
ritonavir with CAP Adp 0.85α 4.35 (pH 6.8), 1.25 (pH 3.8); growth stopped at coverage 0.23σ 0.99[ilevbare2012b R16]
paclitaxel with HPMCAScritical radius 5–7 nm at S 88, smaller than the site spacing of 10–20 nm, so seed growth is not preventedseeded ASD dissolution[que2018 C38]
HPMCAS on felodipinel 68.5 and 15.8 nm; ρc 3.98 nmS 4[schram2016 R12, R14]
posaconazole plates in aspirated human fluidfinger-like ends; step pinning "may be occurring"SEM, 37 °C[elkhabaz2021 R23, C39]
telaprevir crystals with HPMCAS-MFrough surfaces, related to HPMCAS step pinning on felodipine (cited)SEM[lu2018 R10, C48]
Pitfalls and disagreements

The spacing of adsorbed HPMC on felodipine, the central quantity, was not measured (for HPMCAS on felodipine it was imaged by AFM [schram2016 R12]); the statement that molecules sit "relatively far apart" is an inference from α [alonzo2012 C28]. The other mechanisms were excluded by argument [ilevbare2012b C12]. Step pinning applies only below the kinetic roughening transition and was not tested above it [ilevbare2012b X11].

← Concepts and formulas

concept · as used in Crystallization tendency

structural relaxation

The approach of a glass toward the equilibrium supercooled liquid, with loss of excess enthalpy and entropy. In the amorphous solubility papers a phase formed in water is argued to be "much more relaxed" than a melt-quenched glass, with a heat capacity close to that of the supercooled liquid, because water raises molecular mobility and speeds relaxation almeidaesousa2015 C37; telaprevir's glass formed in water is likewise assumed "highly relaxed" mosqueragiraldo2015 C26, C25.

Findings

F2, F7.

Related concepts

concerns the supercooled liquid and glass-liquid phase separation; influenced by plasticization by water; enters the amorphous-crystalline free energy difference.

Formulas

amorphous-crystalline free energy difference from heat capacity.

How it is measured or obtained

Not measured in these papers; inferred from the match between the onset and the supercooled-liquid free-energy estimate, which was lower than the glass estimate for griseofulvin, sorafenib and telaprevir almeidaesousa2015 R6, R11; mosqueragiraldo2015 R13.

Values in the library
materialvalue or classconditionspaper ids
sorafenibonset 0.532 µg/mL; supercooled-liquid estimate 0.479; glass estimate 0.72525 °Calmeidaesousa2015 R6, R11
griseofulvinonset 355.96 µg/mL; supercooled-liquid 356.44; glass 531.4125 °C, HPMCAS-HFalmeidaesousa2015 R5, R11
telaprevironset 90–92 µg/mL; supercooled-liquid Cp 97 ± 8; crystal-to-glass Cp 189 ± 1637 °Cmosqueragiraldo2015 R13
Pitfalls and disagreements

The new phase could not be isolated, so its relaxation state is an inference; the reference state is justified by the agreement it produces almeidaesousa2015 C38, X4; mosqueragiraldo2015 X11.

← Concepts and formulas

concept · as used in Crystallization tendency

sub-Tg nucleation

Nucleation of an amorphous solid in its glassy state, well below Tg, where mobility is low trasi2014b C10. It runs against the rule that storage about 50 °C below Tg prevents crystallization trasi2014b C4.

Findings

F4.

Related concepts

part of nucleation; located by nucleation temperature zone; related to glass transition temperature and polymorphism.

Formulas

none.

How it is measured or obtained

By cooling the melt to low end points and reheating in DSC, by surface nuclei counts after holds below Tg, and by long DSC holds with Raman identification of the product trasi2014b P1, P7, P8.

Values in the library
materialvalue or classconditionspaper ids
nilutamidecrystallization on reheating only after cooling to −30 °C, about 60 °C below TgDSCtrasi2014b R5
nilutamide88% form I after 30 min at −30 °Cabout 50 DSC holdstrasi2014b R16
indomethacin (cited)melting endotherm after storage 55 °C below Tgliteraturetrasi2014b R34
ibuprofen (cited)metastable form nucleates about 60 °C below Tgliteraturetrasi2014b R33
Pitfalls and disagreements

The counts are per sample after 30 min, not rates; longer room-temperature storage nucleated too trasi2014b X7, X16. Mechanical stress or beta relaxation have been proposed as causes elsewhere and were not tested trasi2014b C43, C45.

← Concepts and formulas

concept · as used in Crystallization tendency

supercooled liquid

The amorphous state between Tg and Tm: mobile, but below the melting point; below Tg the same material is a glass lu2016 C2. "Undercooled melt" is used as a synonym baird2012 A1. A glass of a drug–polymer mixture can be regarded as a non-equilibrium supercooled liquid of high viscosity taylor1997 C6. New aspect (C. Amorphous solubility and LLPS): the amorphous reference state of the solubility calculation is the water-saturated supercooled liquid ilevbare2013a C58; almeidaesousa2015 C39, and the drug-rich phase counts as one when its water-plasticized Tg lies below the experimental temperature ilevbare2013a C22; indulkar2015 C38. Ibuprofen, whose Tg of −45 °C makes its drug-rich phase a supercooled liquid at 37 °C, can be treated as an emulsion and seen by solution NMR [ueda2020b C38, M1, C7]. New aspects (Miscibility and drug-polymer interactions): The growth studies measure crystal growth into the supercooled liquid of drug or drug–polymer mixtures, 27–67 °C above the Tg of felodipine (70–110 °C against 43.3 °C), 3–85 °C above that of bifonazole (18–100 °C against 15.2 °C) and up to 61 °C above that of nimesulide (18–80 °C against 19.5 °C) (synthesis: temperature minus Tg) kestur2010a P4, M1; kestur2011 P3, M1, M2. Its lowest point, 18 °C, is at or just below the nimesulide onset Tg kestur2011 X13. Below Tg a binary drug–polymer glass can be regarded as a non-equilibrium, highly viscous supercooled liquid of "solvent" and "solute" molecules with homo- and hetero-associations, which makes the "solution" picture of the indomethacin–PVP dispersion reasonable "at least in terms of molecular interactions if not kinetics" taylor1997 C6, C41, C42. New aspect (Physical stability): in the physical-stability growth studies the supercooled liquid is the melt film, quenched and held between Tg and Tm, from which seeded crystals grow; the polymer effect on growth is placed in this liquid rather than at the crystal faces kestur2013a C1, C6. New aspect (Release mechanism and LoC): whether the drug-rich phase is a supercooled liquid or a glass at the release temperature is offered as a drug property that could matter for the limit of congruency. Que and co-workers (2019) name it "One interesting difference" between ritonavir (supercooled liquid under release conditions, LoC 25%) and ledipasvir (a glass, LoC 5%), and state that further investigation is "clearly" needed [que2019 C49, R28]. For ritonavir Indulkar and co-workers (2019) add that drug-rich nanodomains "can" coalesce into contiguous regions because the drug is a supercooled liquid at 37 °C [indulkar2019a C40]. Saboo and co-workers (2020) place the water-saturated indomethacin-rich phase in the supercooled-liquid state at 25–57 °C and in the glassy state at 10 °C, and find most PVPVA drugs studied "most likely" supercooled liquids at 37 °C in water (all but ledipasvir) saboo2020 C32, C58; at 5 °C loratadine behaves as a high-Tg compound [yang2021 C16].

Findings

Crystallization tendency: F1, F7. C. Amorphous solubility and LLPS: F2, F7, F15, F22. Miscibility and drug-polymer interactions: F1, F10, F13. Physical stability: F3, F18, F19, F27, F28. Release mechanism and LoC: F7, F9, F18.

Related concepts

bounded by glass transition temperature; property melt viscosity; process cold crystallization and crystallization from the supercooled melt; related to crystal growth rate, structural relaxation and plasticization by water; medium for crystal growth inhibition by polymers; related to amorphous solid dispersion; site of glass-crystal growth mode near Tg; related to kinetic versus thermodynamic control of crystal growth. Set by plasticization by water and the glass transition temperature; allows a contiguous drug-rich surface layer to form.

Formulas

Vogel-Tammann-Fulcher equation, Jackson interface-kinetics growth rate, amorphous-crystalline free energy difference from heat capacity; viscosity-limited normal growth law.

How it is measured or obtained

Made by melting and cooling (quench or controlled), then studied isothermally above Tg on a hot stage or in a rheometer lu2016 P1, P2; baird2012 P1. In solution work, a drug-rich phase is a supercooled liquid when its water-plasticized Tg lies below the experimental temperature ilevbare2013a C22; indulkar2015 C38. The drug-rich phase is examined by DSC of the isolated phase, and the free energy of conversion from the crystal to the supercooled liquid is obtained by the Hoffman or heat-capacity cycle ilevbare2013a P9; almeidaesousa2015 E9, E10, E11. In the kestur2010a and kestur2011 growth studies, films of 30–50 µm were made by melting 3–5 mg between cover glasses and cooling rapidly to about 50 K below Tg, with the absence of birefringence checked under crossed polarizers, then nucleated at 40 °C or at ambient temperature and grown on a hot stage kestur2010a P2, P3, P4; kestur2011 P2, R1. Melt-quenched films between cover slips, held on a hot stage, in the physical-stability growth papers trasi2012a P2; trasi2012b P2; mosqueragiraldo2014 P3; kestur2013a P1. In the release-mechanism papers: from the dry or wet Tg against the test temperature, cited or measured [que2019 R28, M1].

Values in the library
materialvalue or classconditionspaper ids
vemurafenibsupercooled liquid between Tg 108 °C and Tm 271 °Cmelt-quenched glasslu2016 M1, R19.a
34 drugsviscosities at Tm from 2.86 × 10⁻³ to 1.99 Pa·srheometrybaird2012 R1
felodipine with 3% polymersgrowth from the supercooled liquid substantially reducedhot stagekestur2010a C28
bifonazole, nimesulidegrowth strongly temperature dependentsupercooled liquidkestur2011 C6
ritonavir drug-rich phaseTg 0.4 °C, so a supercooled liquid at 37 °Caqueous LLPSilevbare2013a C22
clotrimazole; atazanavir precipitatesTg ~14 °C (supercooled liquid at 37 °C); ~51 °C (glass)water at 37 °Cindulkar2015 R22, C38
nicardipine drug-rich phasesupercooled liquid at 37 °C (dry Tg 15 °C); more viscous after 4 h at 2 °Cwaterindulkar2015 M3, R23
griseofulvin; sorafenib new phaseonset matches the supercooled-liquid estimate although the dry drugs are glasses at 25 °C25 °Calmeidaesousa2015 R11, C36
telaprevircrystal-to-supercooled-liquid Cp estimate 97 ± 8 µg/mL against 90–92 µg/mL onset37 °Cmosqueragiraldo2015 R13
felodipinegrowth into the supercooled liquid peaks near 105 °C; 70–110 °C studiedform II, hot stagekestur2010a R5, C10
bifonazolegrowth falls by about 6.4 orders from 100 to 18 °C (plot)melt-quenched filmskestur2011 R7, X6
nimesulidegrowth falls by about 5.4 orders from 80 to 18 °C (plot)melt-quenched filmskestur2011 R9, X6
indomethacin–PVP glassnon-equilibrium supercooled liquid; single Tg citedsolvent-evaporatedtaylor1997 C6, R17
acetaminophenTg 21.2 °C, Tm 169 °CDSC, 10 °C/mintrasi2012a R9.a
flutamideTg −3.1 °C; Tm 108–110 °C with 10 mol% additivesDSCtrasi2012b R7
celecoxibTg 55.34 °C, Tm 163.46 °CDSCmosqueragiraldo2014 R5.a
ATP enantiomers (Tg 16–17 °C)growth from the supercooled liquid at 40–90 °Cmelt films[sato2015 R1, R5]
six compounds (Tg −1 to 90 °C) with PHBgrowth from the supercooled liquid at 30–150 °Cmelt filmssato2016 A4, M1, M2, M3, M4, M5, M6
ritonavirwet Tg around 0 °C; supercooled liquid at 37 °Crelease conditions[que2019 R28; indulkar2019a M1]
ledipasvirglass under release conditions37 °C[que2019 C49, M1]
indomethacin-rich phase (70% dispersion)supercooled liquid at 25–57 °C, slightly below its onset at 17 °C, glass at 10 °Cwet Tg onset 18 °Csaboo2020 C32, R30
loratadinesupercooled liquid at 37 °C, glass at 5 °Cwet Tg 13.8 °C[yang2021 C16, M1]
Pitfalls and disagreements

Near Tg diffusion decouples from viscosity, so viscosity-based growth laws fail there trasi2014a C21. Polymer effects on growth from the supercooled liquid act on the liquid phase rather than at the crystal surface for felodipine kestur2013a C6. In the amorphous solubility papers a glass formed in water may behave as a supercooled liquid, which is inferred and not measured almeidaesousa2015 C37, C38. The text orders of magnitude (5.5 and 6 for bifonazole) are smaller than the plot readings (6.4 and 6.9) kestur2011 X6. Whether dilute dispersions made by cryomilling are molecularly mixed in the melt is assumed kestur2010a C5. In the acetaminophen and flutamide growth papers, interactions in the liquid were judged from spin-coated films at other compositions, not from the growth samples trasi2012b X9; trasi2012a X15. Two drugs only; the idea was not tested [que2019 C49].

← Concepts and formulas

concept · as used in Crystallization tendency

supersaturation

A solution concentration above the crystalline solubility of the drug [almeidaesousa2016 C2; ilevbare2013a C21]. In this chapter the highest supersaturation a solution can hold without crystals is set by the amorphous solubility: above it the excess forms a drug-rich phase, and the concentration and activity of the continuous phase stay at that ceiling mosqueragiraldo2015 C34, C35. Taylor and Zhang 2016 (p. 10) add that the strict upper limit is the spinodal point: a one-phase solution can persist for a time between the binodal and the spinodal, and after phase separation the level is the binodal, which is the amorphous solubility (synthesis). Supersaturation can be generated by solvent shift, pH shift, or dissolution of a high-energy solid form (amorphous drug, dispersion, salt, co-crystal) [almeidaesousa2015 C4, C5; almeidaesousa2016 C2]. The papers of 2017–2022 add that in a micellar medium the supersaturation that matters is the activity ratio a/a^{*} and not the concentration ratio, the two differing about twofold for atazanavir in SDS at the amorphous solubility (65–70 against 33–38) [indulkar2017a R13, C21], and that after a pH shift of a weak-base salt the apparent concentration is held near the amorphous solubility if the dose exceeds it [sugihara2018 R8, R12]. New aspect (Bioperformance): a dissolved drug concentration above the crystalline solubility, with a solute activity above that of a solution saturated with the crystal; the supersaturation ratio is activity over crystal activity, and equals the concentration ratio c/c^* only if the activity coefficients are equal [raina2015c C7, E5, E6, C32]. The highest supersaturation a solution can hold as one phase is set by the amorphous solubility; above it LLPS or crystallization takes over raina2014a C27, C30. Its highest concentration-based value is the ratio of amorphous to crystalline solubility, and that ratio falls when a polymer, a second drug or micelles enter the drug-rich phase or the medium (F18, F19, F20) [ueda2020a E1, C17; alkalla2026 C1, C37; elkhabaz2019b C21, C25]. New aspect (Crystallization from solution): in the crystallization papers supersaturation is the ratio S = C/C_{eq} of the concentration to the crystalline solubility (concentration used in place of activity because the solutions are very dilute), with \sigma = \ln S for growth analyses; the amorphous solubility of ritonavir is S 15.3–15.4 and σ 2.7 [ilevbare2013b E4, C14; ilevbare2012b E1, E2, R3]. The reference solubility is that of the stable crystal, measured without methanol and, to be consistent, with the polymer present (F19) hsieh2012 P4; ilevbare2013c P9, X16; hsieh2014 C18. Other scales in these papers: C/Ceq in the medium of interest (1.5–15), S' = C/Cs with the solubility with additive, the activity ratio δ from membrane flux (2.4–28.8), S 1.5 for acetaminophen, and a nominal 10 mg/mL solvent shift [chen2015 E2; li2016a E1; lu2017 E1; trasi2015 P2; raina2015b P1].

Findings

Amorphous solubility and LLPS: F1, F5, F6, F8, F13, F14, F9, F10, F15, F16, F17, F19, F23. Bioperformance: F3, F10, F13, F14, F18, F19, F20, F28, F29. Crystallization from solution: F1, F2, F9, F14, F19, F21, F22, F27, F31, F34.

Related concepts

capped by amorphous solubility; measured against crystalline solubility; expressed as supersaturation ratio or via thermodynamic activity; relieved by crystallization. Papers of 2017–2022: activity-based supersaturation, maximum achievable supersaturation. Lost by desupersaturation; contrasts with solubilization; summarized by supersaturation factor.

Formulas

supersaturation ratio. Activity-based supersaturation ratio, supersaturation from membrane flux ratio, supersaturation factor, power-law crystal growth rate.

How it is measured or obtained

As concentration divided by the crystalline solubility in the same medium (the supersaturation ratio), or in terms of activity from the flux ratio; the concentration-based ratio at the ceiling is the amorphous-to-crystalline ratio (F6) almeidaesousa2015 E15; mosqueragiraldo2015 R12. In the Bioperformance papers: concentration ratio by UV or HPLC; activity-based ratio from membrane flux relative to the flux at the crystalline solubility [raina2015c E9, R13]; receiver-derived "effective" supersaturation from a Cmax calibration [hate2017 C49, R18]; generated by solvent shift, pH shift or ASD dissolution [raina2014a P7; hate2017 P7; hate2018 P9]. The activity-based maximum is the ratio of the flux from a solution at the amorphous solubility to the flux from a crystalline suspension in the same medium, J_{amph}/J_{eq} [elkhabaz2019b E2, R8, R9]. A dose is described by its ratio to the crystalline solubility: 5 mg of tacrolimus in 100 mL gives S = 20 and in 240 mL S = 8 [hate2019 R7, R14, E2]; pretomanid nucleation is compared at S = 5 nguyen2023a E1, R5. In the crystallization-from-solution papers: concentration by fibre-optic UV on second-derivative spectra or by HPLC; S from the crystalline solubility measured in the same buffer (48–72 h at 37 or 25 °C); the supersaturation factor integrates S over time [ilevbare2012a E2; ilevbare2013c E8].

Values in the library
materialvalue or classconditionspaper ids
ibuprofen to sorafenibmaximum ratio 4.3 to 54.025 °C, fluorescence onsetalmeidaesousa2015 R5
telaprevirmaximum ratio about 18 (stated); flux gain about 18pH 6.8, 37 °Cmosqueragiraldo2015 R14, R15
clotrimazole; atazanavirabout 13; about 6337 °Cindulkar2015 R10
CRH1about 10pH 6.8 and pH 4.6[almeidaesousa2016 R24]
atazanavir in 0–3.6 mg/mL SDSconcentration-based 64.8 → 33.0; activity-based 66.0 → 65.0 at the amorphous solubility37 °C[indulkar2017a R13]
telaprevir in bile saltsconcentration-based value 3.9–6.7 at an activity-based 5.050 mM pH 6.5, 37 °C[lu2017b R17]
felodipine, 25 °C2.7–6.8 at 2–5 µg/mL; about 8 at the amorphous solubilitybufferraina2014a R17
felodipine with BSAS 1.3 and 1.1 by concentration ratio; 7.3 and 4.2 from flux50 µg/mL[raina2015c R13]
nevirapinenominal S 3.9; receiver-derived 2.8 (powder), 4.10 (powder + HPMC-AS)pH shift, 128 µg/mL crystalline[hate2017 R18]
ritonavir (mesoporous-silica study)S 7.8; 3.93.6 µg/mL crystalline[dening2018 E6]
atazanavir, six mediaactivity-based maximum about 50–56; concentration-based 56 in buffer to 14 in pooled human intestinal fluid37 °C, side-by-side cell[elkhabaz2019b R8]
posaconazole, six mediaactivity-based maximum about 29–37; concentration-based about 35–37 in buffer to 5 in human fluid37 °C[elkhabaz2019b R9]
tacrolimusS 20 (5 mg in 100 mL) and S 8 (240 mL); maximum ratio about 33solvent-shift solutions, hollow-fibre module[hate2019 R7, R14; purohit2019 C30]
tacrolimus, fully crystalline capsule contentsconcentration 2.5 times the crystalline solubility during 2 h5 mg in 100 mL, pH 4.5[purohit2019 R5]
atazanavir with darunaviratazanavir in the drug-lean phase about 61–63 instead of 91–96 µg/mL150 µg/mL atazanavir, about 196 µg/mL darunavir added[alkalla2026 R11, C1]
felodipineS 3–12; amorphous benchmark about 10solubility about 0.5 µg/mL, 25 °C[alonzo2012 E1, C42]
ritonavirS 3.8, 7.6 (7.7), 15.3 (15.4) at 5, 10, 20 µg/mL; 4.3 in 50 mM buffersolubility 1.3 µg/mL (100 mM), 2.3 µg/mL (50 mM), 37 °C[ilevbare2012a R3; ilevbare2012b R3]
celecoxib; efavirenzS 6.6, 14.7 at 10 and 22 µg/mL; 1.2 at 10 µg/mLsolubility 1.5 and 8.2 µg/mLilevbare2013b R7
type 1 weak basespeak 1.4–26.7 times the titration solubility, lasting 3.1–14.7 minpH shift, 37 °Chsieh2012 R5
type 2 weak bases7.2–20.7 times the crystalline solubilitypH shifthsieh2012 R10
telaprevir with six bile saltsδ 11.5 and 28.8 need 60 and 150 µg/mL in buffer, 140 and 300–330 with 12 mM STDC/SGDCpH 6.5, 37 °Clu2017 R1
atazanavir, ritonavir in buffer, FaSSIF-V1, Composite-SIFflux-based supersaturation at the amorphous solubility 63–65 and 9.2–10pH 6.5, 37 °C[indulkar2018 R12]
posaconazoleamorphous/crystalline about 34 from pH 2 to 7phosphate buffers, 37 °Cduong2022b R1, C19
Pitfalls and disagreements

The ratio depends on the crystal form (a hydrate for paclitaxel) and the medium [trasi2015a X12]. A solubilizer raises total concentration without raising supersaturation in activity terms, and the concentration-based ratio can fall [ilevbare2013a C54; raina2015c C29]. Total UV concentration is not free drug once a drug-rich phase exists (F4) [almeidaesousa2016 R12]. Concentration ratios understate supersaturation where micelles solubilize the drug, and flux-based values assume that micelles do not cross the membrane [indulkar2017a C42, X9; lu2017b X8]. In the Bioperformance papers, with solubilizers the concentration ratio under-reads the activity-based supersaturation (F3) [raina2015c C38]. Concentration ratios are not valid above the amorphous solubility, where two phases coexist [dening2018 E6]. UV readings above the amorphous solubility include scattered light [hate2018 X15]. In six media of rising solubilizing capacity the concentration ratio fell several-fold while the flux ratio stayed about the same (F18) [elkhabaz2019b R8, R9, C21]. Which dose-to-volume ratio a test imposes decides whether absorption or growth on seeds dominates: S of 8 and S of 20 gave different results in the same module (F28, F29) [hate2019 R14, R17]. Polymer in the drug-rich phase lowers the achievable supersaturation although it inhibits crystallization (F19) [ueda2020a C37]. S for 10 µg/mL ritonavir is printed as 7.6 and 7.7, and for 20 µg/mL as 15.3 and 15.4, depending on rounding (10/1.3 = 7.69; 20/1.3 = 15.38) [ilevbare2012a X1]. The efavirenz induction-time runs state S 2.4, the ratio of the calculated amorphous solubility to the crystalline solubility, while the concentration used gives 2.3 ilevbare2013c X4. A polymer that raises the solubility lowers S for the same concentration (F19) ilevbare2013c X7. C/Cs understates the supersaturation of celecoxib in media where the mass flow does not change [li2016a X18].

← Concepts and formulas

concept · as used in Crystallization tendency

surface crystallization

Nucleation and growth of crystals at a free surface of an amorphous solid, reported to be faster than in the bulk by orders of magnitude because surface molecules are more mobile duong2022c C2; trasi2014a C4. It is a failure mechanism of amorphous solid dispersions in storage and during dissolution duong2022c C2; saboo2020 C36. New aspect (ASD process): in the coating papers the surface route is the target of the treatment, since crystal growth at the free surface of an amorphous solid is orders of magnitude faster than in the bulk, so a thin shell is expected to act where crystals start duong2022c C2, C3; moseson2022a C6. Uncoated powders stored open at 40 °C showed crystals by SEM within 1–2 weeks (ezetimibe–HPMCAS 12–19 days), coated ones later or not at all moseson2022a R11; duong2022c R16, R17. In a coated particle that did crystallize, crystals lay above an intact coating, "as is common for amorphous materials" moseson2022a C26, R15. In dissolution, a hydrated, drug-rich surface layer of bicalutamide–PVPVA crystallizes on the compact surface within 30–60 min (see drug-rich surface layer) moseson2022b R14, R15. New aspect (Supersaturation from ASD dissolution): a proposal for wetted amorphous particles: crystallization "probably" starts at the particle surface, is made worse by water plasticization, and dissolved polymer delays it by interacting with that surface, as a thin polymer film does for amorphous indomethacin (cited) ("reasonable to speculate") alonzo2010 C36, R22, R23. New aspect (Release mechanism and LoC): for indomethacin methyl ester–PVPVA, congruency was lost at a later time point above 5% loading: drug release ceased and polymer release slowed, which the authors attribute to drug crystallization at the compact surface, supported by XRD and PLM saboo2020 C17, R8, R9. A congruency read-out over the first minutes therefore misses this failure. A surfactant built into the dispersion can bring the same failure to a drug that does not crystallize on the binary tablet: ritonavir–PVPVA at 30% with SDS released fast and congruently, then stopped near 80% when needle crystals covered the tablet surface by 15 min, which "likely served as a barrier" indulkar2022 R5, R15, R16, C38. With 10 parts of SDS in ritonavir–PVPVA, needle crystals covered the 30% compact surface at 30 min, to which the authors attribute the release plateau ("can be attributed"); SDS covered about 98 wt% of the dissolved surface, which "conceivably" promotes crystallization through higher surface mobility yang2023 R16, C24, R17, C50.

Findings

Crystallization tendency: F4, F21. ASD process: F43, F44, F38, F40. Supersaturation from ASD dissolution: F1, F2, F14, F24, F27. Release mechanism and LoC: F19, F31.

Related concepts

part of physical stability; related to crystal growth rate and nucleation; contrasts with bulk crystallization in crystallization from the supercooled melt. Targeted by the atomic layer coating through surface molecular mobility; sped up by plasticization by water; in dissolution follows the drug-rich surface layer and leads to matrix crystallization; related to crystallization inhibition by polymers. Ends congruent release after a delay; a form of crystallization during dissolution.

Formulas

none.

How it is measured or obtained

Growth on films with the top coverslip removed trasi2014a P2; nuclei counted on an exposed glass surface trasi2014b P7; SEM, SHG or PLM of particle and tablet surfaces in storage or dissolution duong2022c R16; correasoto2017 R16; saboo2020 R9. SEM was the primary detector of surface crystals on stored coated powder, with XRPD, DSC and SEM agreeing for ezetimibe and XRPD missing several arms of the four-system study moseson2022a C22, X14; duong2022c R16, R18, R20; SEM and SEM-EDX of exposed compacts moseson2022b P14, P15. In the ASD dissolution papers: by polarized light of suspended particles over days [chen2015a P10, R20] and indirectly by the lag in slurry crystallinity with polymer in the medium alonzo2010 R6. In the release-mechanism papers: SEM of the tablet surface at 5, 10 and 20 min, besides XRD, PLM and the 1607 cm⁻¹ IR band of crystalline ester saboo2020 R22, R18, C23. SEM of dried tablets and ATR-IR of blotted wet tablets against the crystalline drug indulkar2022 P9, P11.

Values in the library
materialvalue or classconditionspaper ids
felodipine form Iabout 6 times faster at the surface than in the bulk; about 23 times with 3% PVP30 °C, 0% RHkestur2013b R12
felodipine≈0.06–0.42 µm/s at the surfaceT/Tg ≈1.04–1.11trasi2014a R4.g
loratadine≈0.00005 µm/s at the surface, slowest of 14near Tgtrasi2014a R4.n
nilutamidemuch higher nucleation at the surface than in the bulkmelt-cooled filmstrasi2014b R1
ezetimibe spray-dried dispersionscrystals after 19 days (50% DL) and 12 days (70% DL) uncoated40 °C/75% RHduong2022c C33
uncoated amorphous powders (erlotinib, naproxen and lumefantrine ASDs)crystallization onset within 1–2 weeks40 °C, open dish; 31–75% RHmoseson2022a R11
flutamide spray-dried dispersionneedle crystals growing from particle surfaces40 °C/75% RHcorreasoto2017 C26
indomethacin methyl ester compactssurface crystals within 10 min of dissolutionPVPVA, 25% DLsaboo2020 R9
ritonavir–PVPVA–SDS compactsneedle-shaped surface crystals during dissolution30% DLindulkar2022 R15; yang2023 R16
delamanid dispersion filmsrapid surface crystallization at pH 3.01 hnguyen2023b R3, C37
ezetimibe–HPMCAS 50% DL, uncoatedneedle-like crystals at 19 days; about 26% crystalline at 270 days40 °C/75% RHduong2022c R16, R19
ezetimibe–HPMCAS 70% DL, uncoatedplate-like crystals at 12 days; about 50% crystalline at 270 dayssameduong2022c R16, R19
ezetimibe–HPMCAS coated (50% thin and thick, 70% thick; about 12–40 nm)no crystals by SEM, PXRD or DSC for 24 monthssameduong2022c R16, R17, R18, R20
ezetimibe–HPMCAS 70% thin coating (11.7–15.0 nm)first crystals at 7 months; about 1.5% at 270 dayssameduong2022c R17, R19
erlotinib, naproxen and lumefantrine ASD powders, 13 coated arms3–24 weeks, or none within 48 weeks in 7 arms40 °C, 31–75% RHmoseson2022a R11
extruded and milled 30% and 50% DL BCL–PVPVA compactsneedle-like crystals on the surface at 30 min; a 5–10 µm crystal crust on the 50% compact at 1 hpH 6.8, 37 °Cmoseson2022b R14, R15
amorphous felodipine, polymer in the mediumlag of about 13–20 min before 50% crystallinitypH 6.8, 37 °C slurryalonzo2010 R6
celecoxib–HPMCAS spray-dried particles, Triton X100birefringent regions on or in the particles after about 7 days, no free needlespH 3, 25 °C[chen2015a R20, C37]
same, no surfactantcrystallization only after more than 7 days, apparently from matrix or surfacepH 3[chen2015a C33]
delamanid salt dispersions, five enteric gradescrystals on films and powders after 1 h in acid, more with HPMCAS and at higher pH; HPMCAS-HF also at pH 1.625% drug, 37 °C[nguyen2023c R19, R18]
delamanid dispersions after the pH shiftundissolved particles covered with small crystals within 30 minpH 6.5[nguyen2023c R21]
indomethacin methyl ester–PVPVA 25%smooth at 5 min; scattered needle-like crystals at 10 min; surface extensively covered at 20 minSEM, pH 6.8saboo2020 R22, M6
indomethacin methyl ester–PVPVA 10–25%drug release levelled off after 7.5–20 min30 min runssaboo2020 R8
ritonavir–PVPVA 30:70:5 with SDSsmooth at 2 min; needle crystals and crystalline-ritonavir IR bands at 15 min; release flat near 80%SEM, ATR-IR, pH 6.8indulkar2022 R15, R16, R5
ritonavir–PVPVA–SDS 30–70–10birefringent needles at 30 minPLM, SEM of the compact surfaceyang2023 R16
Pitfalls and disagreements

Screens with films sandwiched between cover glasses assume surface crystallization is minimized, which was not tested baird2010 C45. Surface and bulk data are often taken at different temperatures trasi2014a X16. Where surface growth exceeds bulk growth, extrapolating bulk data may overpredict stability kestur2013a C18. The coating delays surface crystallization but does not stop it: coated arms failed at 3, 6 and 24 weeks and at 7 months, and the benefit depends on humidity through the wet glass transition, on the polymer and, for a fast crystallizer at high loading, on thickness (see F44) moseson2022a R11, R12; duong2022c R17. Neither surface mobility nor water uptake during storage was measured moseson2022a C4; duong2022c X7. XRPD shows no crystals through 48 weeks for uncoated extruded lumefantrine powders that SEM scored at 1 week moseson2022a X14. Where crystallization starts on the wetted felodipine particles and whether polymer adsorbs there were not measured alonzo2010 C36. The 7-day observations of the celecoxib suspensions appear only in the discussion and are imaged for Triton X100 only [chen2015a X17]. The authors read the crystals as forming mainly at the surface of the wetted matrix, where water lowers the Tg and raises mobility, and as acting as seeds once the polymer dissolves [nguyen2023c C43, C44]. The 5-min rates used to set the limit of congruency did not show the later stop of release saboo2020 X7. Crystals were imaged only at 30% and not identified yang2023 X12.

← Concepts and formulas

concept · as used in Crystallization tendency

surface-enhanced crystal growth

Faster crystal growth at a free surface of an amorphous solid than in its bulk, thought to reflect higher surface mobility near or below Tg trasi2014a C4. In this chapter it is offered as one reason why felodipine growth in uncovered thin films outpaced the bulk mobility predicted from Tg rumondor2010 C28, C29.

Findings

F5, F9, F21, F26.

Related concepts

part of crystal growth rate; related to surface crystallization and molecular mobility.

Formulas

constant-penetration-depth crystallinity estimate.

How it is measured or obtained

Radial growth rates of crystals in films with the top coverslip removed, compared with sandwiched films trasi2014a R4, R5; growth in uncovered spin-coated films at fixed RH rumondor2010 P1, P5.

Values in the library
materialvalue or classconditionspaper ids
amorphous felodipine films (uncovered)growth above the AGV mobility line; coupling exponent 0.513–75% RH, 25 °Crumondor2010 R7, C27, C28
14 compounds (outside)surface growth generally faster than bulk; order differs from bulknear Tgtrasi2014a R4, C4, C8
felodipine form Isurface ≈6 times the bulk at 18 °C; equal above 70 °Cmelt filmskestur2013a R8.a, R9, C2
felodipine form I with 3% PVPsurface/bulk ratio 22.8 against 5.7 without30 °Ckestur2013b R12, C32
felodipine powders, 150–250 against 500–710 µm≈46 against ≈12% crystalline at 24 h30 °C, Ramankestur2013b R3.e, C24

Evidence table: surface-enhanced crystal growth.

Pitfalls and disagreements

In the felodipine growth study surface growth is one of two "possible explanations", next to diffusion–viscosity decoupling; neither surface nor bulk growth was measured separately there rumondor2010 C28, X12. The authors consider thin uncovered films relevant because crystallization "likely" starts at the surface rumondor2010 C30. The constant-depth picture is not consistent with the bulk growth rate with PVP, which would cross the estimated depth within days kestur2013b X8.

← Concepts and formulas

concept · as used in Crystallization tendency

surface molecular mobility

The mobility of molecules at the free surface of an amorphous solid, which is taken to be much higher than in the bulk and to be the reason why crystals nucleate and grow faster at a surface (see surface crystallization) duong2022c C2. In the coating papers it is the mechanism proposed for the protection by an atomic layer coating: the shell lowers surface mobility, "attributed primarily" in the study of four systems and "perhaps more importantly" than a moisture barrier in the ezetimibe study moseson2022a C4, C44; duong2022c C53, C54. Water raises mobility and lowers the wet glass transition temperature, so a coating that acts on the surface cannot rescue a powder whose bulk is mobile moseson2022a C41, C25.

Findings

F43, F44.

Related concepts

reduced by the atomic layer coating; raised by plasticization by water; drives surface crystallization; the alternative explanation is the moisture barrier effect.

Formulas

Tg − T.

How it is measured or obtained

Not measured in any ASD process paper moseson2022a C4; duong2022c X7. It is inferred from crystallization onset in coated and uncoated powders and from the wet Tg minus the storage temperature, computed with the onset Tg after 24 h at the storage humidity (see Tg − T) moseson2022a P6, E2, R11, R12.

Values in the library
materialvalue or classconditionspaper ids
erlotinib–HPMCAS, 75% RHwet Tg − T about −8.8 °C; coated onset about 3 weeks (uncoated about 2)40 °Cmoseson2022a R12, R11
erlotinib–HPMCAS, 43% RH−2.3 ± 1.1 °C; coated none at 48 weeks40 °Cmoseson2022a R12, R11
erlotinib–PVPVA, 43% RH−0.5 ± 1.2 °C; coated onset about 6 weeks40 °Cmoseson2022a R12, R11
naproxen–HPMCAS 75% RH; lumefantrine–PVPVA 43% and 31% RHabout +11.3, +10.7 and +14.3 °C; coated 24 weeks, 24 weeks and none at 48 weeks40 °Cmoseson2022a R12, R11
uncoated powders of all four systemscrystals within about 1–2 weeks whatever Tg − T40 °C, 31–75% RHmoseson2022a R11, X15
Pitfalls and disagreements

The authors write that "this confirms that the mobility of the sample was the primary mechanism" of the coated failures moseson2022a C25; uncoated powders failed in 1–2 weeks across the whole range of Tg − T, the stable erlotinib–HPMCAS arm at −2.3 °C lies outside the range of about 0–15 °C that the discussion gives, and whether the wet Tg was measured on coated or uncoated powder is not stated moseson2022a X15, C45. The ezetimibe study offers both mobility and a moisture barrier and could not separate them duong2022c C53, C54, X7.

← Concepts and formulas

concept · as used in Crystallization tendency

thermodynamic driving force for crystallization

The free-energy difference between the supercooled liquid (amorphous form) and the crystal, written ΔGv per unit volume or ΔG per mole or gram; it is zero at Tm and grows on cooling baird2012 C32; vaneerdenbrugh2010a E2. A larger driving force lowers the classical nucleation barrier vaneerdenbrugh2010a E3, C39. This draft comes from the crystallization-tendency chapter; for drugs in polymers the same quantity is lowered by mixing konno2006 C34. New aspect (PEG dispersions): in a dispersion in a crystalline polymer the driving force of the drug is lowered by melting point depression, because the supercooling at the storage temperature falls. Zhu and co-workers attribute the delayed crystallization of benzocaine in PEG to entropic mixing and favourable benzocaine–PEG interactions that depress its melting point (around 48 °C at the eutectic against around 89 °C for the neat drug), so that supercooling and driving force at 25 °C are much smaller; with a Tg of −31 °C they consider a thermodynamic rather than a kinetic cause "reasonable" zhu2012 C32, C33. New aspects (Miscibility and drug-polymer interactions): In a miscible dispersion the chemical potential of the drug, and so the driving force for crystallization, must be lower than for the pure amorphous drug, through the entropy of mixing and, if adhesive interactions exceed cohesive ones, the enthalpy; melting point depression is its visible sign marsac2006b C38, C39. In the PVP dispersions the predicted reduction is small: the Hoffman estimate "does not forecast a large reduction", especially at lower temperature, and is "most likely underestimated" because equilibrium depression is hard to reach at high polymer content marsac2006b C41, C40. At low polymer levels, where the melting point and Tg of the drug are unchanged, there "does not appear" to be a macroscopic effect on ΔG, so polymer effects on growth are not read as thermodynamic kestur2010a C36, C41. New aspect (Physical stability): for dispersions holding water, the driving force is framed as the degree of supersaturation of the drug in the polymer–water co-solvent, \sigma = (\mu^{1} - \mu^{0})/RT, which water raises as an anti-solvent; it was reasoned, not computed marsac2008a E1, C26, C49, X4.

Findings

Crystallization tendency: F2, F6, F7, F13. PEG dispersions: F8, F9. Miscibility and drug-polymer interactions: F9, F10, F11. Physical stability: F2, F4, F18, F22, F28, F29.

Related concepts

measured by Hoffman equation; depends on enthalpy of fusion, entropy of fusion and degree of supercooling, which is reduced in a dispersion via melting point depression; part of classical nucleation theory; related to nucleation, crystal growth rate and heterogeneous nucleation; lowered by entropy of mixing; contrasts with kinetic stabilization; related to crystal growth inhibition by polymers, supersaturation, cosolvency and kinetic barrier to nucleation; balanced against mobility in kinetic versus thermodynamic control of crystal growth.

Formulas

Hoffman equation, classical nucleation barrier, entropy of fusion, solid-liquid equilibrium solubility equation, Flory-Huggins activity equation.

How it is measured or obtained

Mostly estimated with the Hoffman equation from Tm and ΔHfus vaneerdenbrugh2010a E2; baird2010 E25; lu2016 E1; from MDSC heat capacities, which agreed with the Hoffman estimate for felodipine and nifedipine marsac2006a R11, C14; as \Delta G \approx \Delta S_m\,\Delta T for growth analysis trasi2014a E3; for dispersions, from melting-point depression and Flory–Huggins mixing marsac2006b C41; marsac2009 C34. In the PEG dispersions it was not measured: the melting points are cited and the driving force is inferred from the phase diagram zhu2012 C32, M3. For the PVP dispersions, estimated with the Hoffman equation, for pure nifedipine from 0 to 180 °C and, with the depressed melting temperature, for 30 and 40% PVP marsac2006b P8, E4, R12; for the dispersions the melting temperature of the 40% PVP curve is not shown in the paper's own melting data marsac2006b X5. The unchanged offset melting point of physical mixtures with 3% polymer is taken as an unchanged driving force at a given temperature kestur2010a R1, C36. In the moisture papers: Hoffman estimates from Tm and heat of fusion for six drugs at 25 °C [rumondor2009c E3, M1]; crystal solubility in NMP and NMP–water as stand-ins for PVP and PVP–water marsac2008a P8.

Values in the library
materialvalue or classconditionspaper ids
nifedipineΔG ≈−30, ≈−24, ≈−16 J/g at 0, 50, 100 °CMDSC and Hoffmanmarsac2006a R11
felodipineΔG ≈−18, ≈−13, ≈−7 J/g at 0, 50, 100 °CMDSC and Hoffmanmarsac2006a R11
34 drugsΔGv from −8.6 (lidocaine) to −54.7 J/cm³ (acetaminophen)Hoffman, 298 Kbaird2012 M5, M12
class I against class IIIΔGv ≈−0.40 against ≈+0.57 SD from the meanper volume, 293.15 Kbaird2010 R22.g
spin-coat classes I, II, III≈−0.29, ≈+0.29, ≈+0.13 SDper volumevaneerdenbrugh2010a R20.e
vemurafenib α; β≈−11.0; ≈−6.3 kJ/mol at about 115 °CHoffman, plotted as crystal minus liquidlu2016 R22, X17
nifedipine with 40% PVPreduction smaller than the nifedipine–felodipine differencemelting-point depressionmarsac2006b C41
drugs in PVPdriving force remains at most practical polymer contentsinteraction parametersmarsac2009 C34
benzocaine in PEG 3350melting point depression from around 89 to around 48 °C at the eutecticcitedzhu2012 C32, M3
ibuprofen; fenofibrate in PEG 3350melting point depression of about 30 °C; about 20 °C (text), about 24 °C (plot)eutecticzhu2012 C34, X11
nifedipine, pure; 30% PVP; 40% PVP−9.0, −8.4, −7.95 kJ/mol at 20 °C (zero at ≈171, ≈148, ≈128 °C)Hoffman, plot readingsmarsac2006b R12
nifedipine, pure; 30% PVP; 40% PVP−5.45, −4.1, −2.7 kJ/mol at 100 °Cas abovemarsac2006b R12
felodipine with 3% PVP, PVP/VA, HPMCAS, PVAcmelting point of physical mixtures unchanged (140.7–142.9 °C against 141.5 °C)offset, 1 °C/minkestur2010a R1
bifonazole, nimesulide with 5% polymersTg rise 0.4–2.2 °CDSC onsetkestur2011 R6, C12
nifedipine, felodipine, droperidol, pimozide, indomethacin, ketoprofenHoffman ΔGc 8.2, 6.1, 9.2, 11.0, 8.1, 4.1 kJ/mol25 °C[rumondor2009c M1, M6, M3, M2, M4, M5]
nifedipine vs felodipine crystalsmole-fraction solubility 0.05 vs 0.10 in NMP: larger lattice term for nifedipine25 °Cmarsac2008a R10, C35
nifedipine and felodipine in PVPFlory–Huggins activity ≈0.84 and ≈0.83 at Φ 0.2χ −3.8, −4.2marsac2006a R19
acetaminophen and flutamide with 10% polymerTm barely changed, so the driving force is similar for all polymersDSCtrasi2012a R9, C19; trasi2012b R7, C33
celecoxib with TPGS and PVPTm 12 °C lower; a smaller driving force "could contribute" to slower growth near TmDSC, melt growthmosqueragiraldo2014 R5.f, C33
six compounds with PHBdriving force inferred unchanged from Tm and IR ("can be inferred")DSC; IRsato2016 C18, X11
indomethacin polymorphsdifferences in growth persist after a driving-force normalizationmelt growthtian2017 R11, C24
Pitfalls and disagreements

The class trend is not monotonic: class II, not class III, had the least negative ΔGv in the solvent screen vaneerdenbrugh2010a X10. Pair comparisons of viscosity can be confounded by driving force baird2012 X9. Indomethacin has a larger driving force than felodipine yet crystallizes more slowly during dissolution alonzo2010 C42, and metastable vemurafenib β outgrows α despite α's larger driving force lu2016 C52. Low polymer levels barely change it, so polymer effects on growth are read as kinetic kestur2010b C28; thermodynamically stable dispersions are hard to achieve marsac2009 C34. For ibuprofen and fenofibrate in PEG the depression should lower the driving force, yet the onset in the dispersion is earlier than for the neat drug; the authors explain it by nucleation on crystalline PEG, not by the driving force zhu2012 C34, C41. The estimated solubilities of benzocaine (28%) and ibuprofen (25%) exceed the 20% loading, which the card's extractor notes as not discussed zhu2012 X13, M3, M4. The 40% PVP curve rests on a melting temperature not shown among the measured data marsac2006b X5. The unchanged-driving-force conclusion for 3% polymer uses the melting point of form I in physical mixtures while form II grows kestur2010a X7. The size of the lowering differs between the polymers (synthesis: contrast between papers): in PEG 3350 the melting point depression (benzocaine from around 89 to around 48 °C at the eutectic) is taken to make supercooling and driving force much smaller zhu2012 C32, C33, whereas for PVP the predicted reduction is small marsac2006b C41. The driving force did not sort drugs by moisture-induced phase separation [rumondor2009c C32]. The supersaturation explanation of the nifedipine–felodipine ranking with water rests on NMP data and printed cosolvency fits that do not reproduce their points marsac2008a X2, X4. The celecoxib driving force was argued from Tm and not calculated mosqueragiraldo2014 C33.

← Concepts and formulas

concept · as used in Crystallization tendency

Tm − Tg temperature gap

The span between the melting point and the glass transition, the range a melt must cross to become a glass trasi2014a C27. If viscosity at Tg is similar for all compounds, a small Tm − Tg with a high viscosity at Tm means a high viscosity at the temperature of fastest growth and so a low maximum growth rate trasi2014a C27.

Findings

F5, F12.

Related concepts

related to reduced glass transition temperature; related to melt viscosity and crystal growth rate.

Formulas

linear fit of melt viscosity on Tm − Tg.

How it is measured or obtained

From DSC Tm and Tg trasi2014a M1; baird2012 M1.

Values in the library
materialvalue or classconditionspaper ids
ritonavir; tolazamide77; 154 °CDSCbaird2012 M34, M18
chlorpropamide; celecoxib; ketoconazole108; 105; 105 °C, in classes I-B, II, IIIDSCbaird2012 M9, M14, M26
slow and fast growth groups85–117 and 121–146 for drugs with viscosity dataDSCtrasi2014a R9
class means I-A, I-B, II, III123 ± 11, 119 ± 11, 126 ± 20, 108 ± 16 °C, not significantly differentDSCbaird2012 R10
Pitfalls and disagreements

The celecoxib Tm − Tg of 125 in one table conflicts with its own Tm and Tg (108) trasi2014a X1, and the dibucaine row of another table repeats lidocaine baird2012 X1. The gap tracked growth groups but not the DSC classes (synthesis: compare R9 of trasi2014a with R10 of baird2012).

← Concepts and formulas

formula · as used in Crystallization tendency

amorphicity index

Expression

A score for a spin-coated film or a drug–polymer pair, built from the category codes assigned by polarized light microscopy at four time points. Each sample gets a four-letter code (AAAA completely amorphous to CCCC fully crystalline), and the amorphicity index counts its amorphous letters:

\mathrm{AI}\ (\%) = \frac{\text{number of A letters in the four category codes of a sample}}{16}\times 100 vaneerdenbrugh2010b E1

(printed in words; 16 is the number of A letters of a sample that is AAAA at all four time points). The scores are then averaged over the seven drug/polymer weight ratios of a pair, over all drugs for one polymer, and over all polymers for one drug (all printed in words):

\overline{\mathrm{AI}}_{\mathrm{pair}} = \frac{1}{7}\sum_{i=1}^{7}\mathrm{AI}_i vaneerdenbrugh2010b E2;

\overline{\mathrm{AI}}_{\mathrm{polymer}} = \frac{1}{N_{\mathrm{drugs}}}\sum_{\mathrm{drugs}}\overline{\mathrm{AI}}_{\mathrm{pair}} vaneerdenbrugh2010b E3;

\overline{\mathrm{AI}}_{\mathrm{drug}} = \frac{1}{N_{\mathrm{polymers}}}\sum_{\mathrm{polymers}}\overline{\mathrm{AI}}_{\mathrm{pair}} vaneerdenbrugh2010b E4.

The 2011 paper prints the same definition, with the index of a drug–polymer pair as the mean of the indices of its weight ratios [vaneerdenbrugh2011a E1, P3].

Variables and units

symbolmeaningunits
AIamorphicity index of one sample% vaneerdenbrugh2010b E1
A letteramorphous letter in the four-letter category, one for each of day 0, 1, 3 and 7count, 0 to 4 per sample vaneerdenbrugh2010b P2, E1
\overline{\mathrm{AI}}_{\mathrm{pair}}mean AI over the 7 drug/polymer ratios (90/10 to 10/90 w/w)% vaneerdenbrugh2010b E2, P1
N_{drugs}, N_{polymers}8 drugs and 7 polymerscount vaneerdenbrugh2010b R2

Assumptions and valid domain

The categories are semiquantitative and come from the extent of birefringence under polarized light, which is assumed to follow the amount of crystalline drug because the films are thin and crystallization extends over the full film height vaneerdenbrugh2010b C11, P2. The same weight is given to the four time points and to each ratio vaneerdenbrugh2010b E1, E2. Defined for dry-stored spin-coated films from ethanol, or from ethanol/dichloromethane for HPMC and HPMCAS vaneerdenbrugh2010b P1.

Use in the library

paperused forvalues or fitted parametersstatus / recomputed
vaneerdenbrugh2010bamorphicity of 392 spin-coated drug–polymer films (synthesis: 8 drugs × 7 polymers × 7 ratios)BD-E100 25/75 = 44; FFA-PAA mean 13; PVP mean 70; BD mean 48reproduces (4 + 1 + 1 + 1 = 7 A letters, 7/16 = 43.75%; FFA-PAA mean 12.5; PVP mean 70.0; BD mean 47.6) vaneerdenbrugh2010b E1, E2, E3, E4
vaneerdenbrugh2010bper-drug rankingPH 47, CZ 47, BD 48, LI 55, FFA 59, FB 68, CP 73, BI 97derived vaneerdenbrugh2010b R4
vaneerdenbrugh2010bper-polymer rankingPVP 70, PVPVA 70, E100 67, HPMC 66, HPMCAS 62, PAA 49, PSSA 48derived vaneerdenbrugh2010b R5
vaneerdenbrugh2011aamorphicity of 105 spin-coated pairs (15 drugs × 7 polymers; ratios 75/25, 50/50, 25/75 and five more for the drugs of the first screen)PH–PSSA 78; FFA–PAA 13; the 8 repeated drugs equal the first screenreproduces: a sample scored AAAA at all four time points has 4 × 4 = 16 A letters, AI 100%; per-sample categories not printed [vaneerdenbrugh2011a E1, R3, X9]

Where it fails or is disputed

One entry of the supplementary table (FFA–PSSA 60/40, printed 94 where the category codes give 6) does not reproduce, and the averages for FFA–PSSA, FFA and PSSA reproduce only with the printed value vaneerdenbrugh2010b X1. The averages hide the drug-by-drug reversal of the acidic polymers: PAA and PSSA are the best polymers for basic drugs and the worst for acidic drugs, yet their polymer means (49, 48) are the lowest of the seven vaneerdenbrugh2010b R5, C3. Film thickness, total solids concentration and storage temperature are not stated, so scores can be compared with other papers only qualitatively vaneerdenbrugh2010b X10. The cellulosic polymers were cast from a different solvent, which confounds their scores with solvent vaneerdenbrugh2010b X12. The paper reports only averaged scores, not the polymer fraction needed for each drug vaneerdenbrugh2010b X13. In the 2011 screen the number of ratios averaged per drug is not stated, there are no error bars, and one changed letter moves a three-ratio mean by about 2.1% [vaneerdenbrugh2011a X9, X10].

Concepts

crystallization inhibition, crystal growth inhibition by polymers, drug-polymer interactions, crystallization tendency. Findings. F15, F16, F24.

← Concepts and formulas

formula · as used in Crystallization tendency

amorphous-crystalline free energy difference from heat capacity

Expression

With linear reversing heat capacities C_p^{sl} = aT + b and C_p^{c} = cT + d almeidaesousa2015 E6, E7, the crystal-to-supercooled-liquid change at T is

\Delta H_{c\to sl} = \Delta_{fus}H + \frac{aT^2 + 2bT}{2} - \frac{aT_m^2 + 2bT_m}{2} + \frac{cT_m^2 + 2dT_m}{2} - \frac{cT^2 + 2dT}{2} almeidaesousa2015 E9,

\Delta S_{c\to sl} = \frac{\Delta_{fus}H}{T_m} + (T_m - T)(c - a) + \ln\left(\frac{T_m}{T}\right)(d - b) almeidaesousa2015 E10,

\Delta G_{c\to sl} = \Delta H_{c\to sl} - T\,\Delta S_{c\to sl} almeidaesousa2015 E11.

A glass version uses C_p^{g} = eT + f below Tg almeidaesousa2015 E8, E12, E13, E14. The integral form printed for telaprevir is

\frac{\Delta G_{a\to c}}{RT} = \frac{\Delta H_f}{RT}\left(1 - \frac{T}{T_m}\right) + \frac{1}{RT}\int_{T_m}^{T}\Delta C_p\,dT - \frac{1}{R}\int_{T_m}^{T}\frac{\Delta C_p}{T}\,dT mosqueragiraldo2015 E3.

Variables and units

symbolmeaningunits
a, b; c, d; e, flinear fit parameters of the reversing Cp of supercooled liquid, crystal, glassJ mol⁻¹ K⁻¹ and J mol⁻¹ K⁻² almeidaesousa2015 E6, E7, E8
\Delta_{fus}H, T_menthalpy of fusion, melting temperaturekJ/mol, K almeidaesousa2015 E9
\Delta H, \Delta S, \Delta Gconversion enthalpy, entropy, free energy at TkJ/mol, J mol⁻¹ K⁻¹, kJ/mol almeidaesousa2015 R12

Assumptions and valid domain

Cp linear in temperature over the range of interest (shown for tolnaftate only); un-ionized, non-solvated crystal that melts without degradation; only the reversing Cp is used because the entropy change is the reversibly exchanged heat divided by T almeidaesousa2015 C10, C11, E17. For much larger undercooling the form with a 1/T^2 term is better almeidaesousa2015 E5, C10.

Use in the library

paperused forvalues or fitted parametersstatus / recomputed
almeidaesousa2015six drugs at 25 °C; sorafenib and griseofulvin also to the glassΔG 3.56–10.34 kJ/mol (to supercooled liquid); 11.37 and 10.38 kJ/mol (to glass)ΔG = ΔH − TΔS reproduces from Table 7 (9.39 kJ/mol for griseofulvin); parameters a–f not printed almeidaesousa2015 E11, R11, R12
mosqueragiraldo2015telaprevir at 37 °C97 ± 8 and 189 ± 16 µg/mL through Eq. 1ΔCp values not printed mosqueragiraldo2015 E3, R13
almeidaesousa2016CRH15.85 and 5.95 kJ/molmethod named only, parameters not printed [almeidaesousa2016 E5, M1]

Where it fails or is disputed

The method's superiority is concluded on the hypothesis that the onset is governed by the supercooled liquid, and neither ΔG nor the amorphous solubility was determined independently almeidaesousa2015 X4. Eq. 4 is printed with a symbol clash with Eqs. 5–7 and is not used almeidaesousa2015 X13.

Concepts

amorphous-crystalline free energy difference, supercooled liquid, structural relaxation. Findings. F7.

← Concepts and formulas

formula · as used in Crystallization tendency

amorphous solubility advantage from free energy difference

Expression

\ln x_{SL,L_1} = \ln x_{C,L_1} + \frac{\Delta G_{a\to c}}{RT} ilevbare2013a E5;

\frac{\mu_1^{\circ} - \mu_{1c}^{\circ}}{RT} = \ln\frac{S_a}{S_c} [trasi2015a E1];

\frac{S_{amorphous}}{S_{crystalline}} \approx \frac{a_{amorphous}}{a_{crystalline}} = \exp\left[\frac{\Delta G_c}{RT}\right] hsieh2012 E4; hsieh2014 E3.

The ASD dissolution papers print it as

\Delta G_c = RT\ln\frac{a_{\mathrm{amorphous}}}{a_{\mathrm{crystalline}}} \approx RT\ln\frac{S_{\mathrm{amorphous}}}{S_{\mathrm{crystalline}}} [konno2008a E1];

\frac{\sigma^{\mathrm{amorph}}}{\sigma^{\mathrm{crystal}}} = e^{\Delta G/RT} [alonzo2011 E2];

\frac{S_A}{S_C} = \exp\left(\frac{\mu_A - \mu_C}{RT}\right) [li2018 E2];

\Delta G = -RT\ln\left(\frac{C_a}{C_s}\right) duong2022a E1.

Variables and units

symbolmeaningunits
x_{SL,L_1}, x_{C,L_1}mole-fraction solubility of the supercooled liquid and of the crystal in waterdimensionless ilevbare2013a E3, E4
\Delta G_{a\to c}free-energy difference amorphous–crystalilevbare2013a E2
\mu_1^{\circ}, \mu_{1c}^{\circ}chemical potential of pure amorphous (supercooled liquid) and crystalline drug[trasi2015a E1]
S_a, S_camorphous and crystalline solubilityµg/mL [trasi2015a E1, R1]
S_{amorphous}, S_{crystalline}solubility of the amorphous and crystalline formsµg/mL hsieh2012 E4
a_{amorphous}, a_{crystalline}thermodynamic activity of the solid formdimensionless hsieh2012 E4
\Delta G_cfree-energy difference between the forms (Hoffman equation)kJ/mol hsieh2014 E3
R, Tgas constant; temperatureJ mol⁻¹ K⁻¹; K
\sigma^{\mathrm{amorph}}, \sigma^{\mathrm{crystal}}solubility of each form (2011 notation)µg/mL [alonzo2011 E2]
S_A, S_Camorphous and crystalline solubility (2018 notation)µg/mL [li2018 E2]
\mu_A, \mu_Cchemical potential of the drug in the amorphous and the crystalline solidJ/mol [li2018 E2]
\Delta G; C_a, C_sfree-energy difference; amorphous and crystalline solubility (2022 notation)J/mol; µg/mL duong2022a E1

Assumptions and valid domain

The activity coefficient is constant between the crystalline and amorphous concentration; the amorphous phase is a supercooled liquid; the first form omits the water term, which amorphous solubility advantage with water-sorption correction adds [ilevbare2013a C55, C58; trasi2015a C36]. In the activity form the ratio of solubilities equals the ratio of activities; water mixing into the amorphous phase is neglected, which can lower the amorphous solubility by up to 2-fold hsieh2014 R17. Used at 25 and 37 °C. In the ASD dissolution papers the activity ratio is approximated by the solubility ratio [konno2008a E1]; solute–solvent interactions are taken as equal for both solid forms, which the 2011 authors call "unlikely to be true" for a dispersion [alonzo2011 C6, C41]. The drug activity coefficient in solution varies little between the crystalline and the amorphous solubility, and the amorphous form and the solution equilibrate; crystallization, water ingress and glass relaxation hamper that equilibrium [li2018 C41].

Use in the library

paperused forvalues or fitted parametersstatus / recomputed
ilevbare2013afirst approximation of the amorphous solubility, ritonavirnone printednot applicable (no numerical output) ilevbare2013a E5
trasi2015adefines the amorphous-to-crystalline solubility ratioritonavir 15–16, lopinavir 6.0–6.4, paclitaxel hydrate 85–94, felodipine 7–8 (extractor arithmetic)not applicable; ratios from Table 1 [trasi2015a E1, X12]
hsieh2012uncorrected amorphous solubility of five basesratios 22.2, 11.9, 20.4, 54.6, 7.0ratios reproduce from printed solubilities (24.4/1.1 = 22.2) hsieh2012 E4, R10
hsieh2014glyburide and warfarinabout 60 and 48reproduces 59.9 and 47.7 at 25 °C hsieh2014 E3, R16
konno2008afelodipine at 37 °Cratio about 10: 10 × 0.94 = 9.4 against printed about 9.5 µg/mLreproduces within rounding [konno2008a E1]
alonzo2011felodipine at 37 °Cratio about 9 implies ΔG = RT ln 9 ≈ 5.67 kJ/molinputs not printed [alonzo2011 E2]
alonzo2011flux of pure amorphous felodipine against crystalratio 8.5close to the estimate [alonzo2011 R14, C40]
li2018lopinavir, pH 3, 25 °Cmeasured S_A/S_C = 17.4/3.1 = 5.6; \mu_A - \mu_C not printedinputs not printed [li2018 E2, R1] (synthesis: RT\ln 5.6 ≈ 4.3 kJ/mol at 298 K)
duong2022aturning a Hoffman free-energy estimate into a delamanid solubility ratiomeasured ratio 42.3 at pH 6.5 against 31.8 estimatedthe two equations are used together; no ΔG value is printed duong2022a E1, E2, R27

Where it fails or is disputed

The paclitaxel ratio lies outside the 5–50-fold range quoted in the introduction and rests on a hydrate [trasi2015a X12, R18]. The ketoconazole estimate (54.6-fold) is 2.6 times the measured supernatant, and the water correction brings it to 15.3-fold hsieh2012 X2, R10.d. For warfarin with Eudragit E100 the polymer-free supersaturation of 60.5 exceeds the 48-fold ceiling hsieh2014 X4. A dispersion and the pure amorphous drug need not give the same solution activity; their measured equality for felodipine "may be purely coincidental" [alonzo2011 C45, C48]. For lopinavir the measured ratio is used; the free-energy difference from thermal data is printed but not applied to data [li2018 E3, C43]. The delamanid ratio was measured in PBS pH 6.5, where both solubilities are very low (0.76 and 0.018 µg/mL), so small absolute errors move the ratio duong2022a R1.

Concepts

amorphous solubility advantage, chemical potential, amorphous solubility, Hoffman equation. Findings. Amorphous solubility and LLPS: F6, F11. Crystallization from solution: F2. Supersaturation from ASD dissolution: F7, F18, F23.

← Concepts and formulas

formula · as used in Crystallization tendency

Arrhenius equation

Expression

For the nucleation rate:

I = A\exp\left(-\frac{\Delta E}{RT}\right) marsac2006a E2

as a slope, for a crystallization rate constant (outside the subcategory):

\frac{d\ln k}{d(1/T)} = -\frac{\Delta E}{R} [wanapun2010 E5]

For melt viscosity:

\log\eta = \log\eta_0 + \frac{E_\eta}{2.303RT} tian2017 E1

Variables and units

symbolmeaningunits
Inucleation rateunit not stated on the plot marsac2006a X2
Apre-exponential factor– marsac2006a E2
\Delta Eactivation energykJ/mol marsac2006a R3
R, Tgas constant; absolute temperatureJ mol⁻¹ K⁻¹; K marsac2006a E2
\eta, \eta_0melt viscosity; constantunit not printed tian2017 E1
E_\etaactivation energy for viscous flowkJ/mol tian2017 E1

Assumptions and valid domain

Fitted separately above and below Tg, since the temperature dependence breaks at Tg marsac2006a P5, C10. For viscosity the relation "may" hold over small temperature ranges, as a straight line of log η against 1/T tian2017 E1.

Use in the library

paperused forvalues or fitted parametersstatus / recomputed
marsac2006aactivation energy of nucleation, pure and with 3 wt% PVPbelow Tg 122 (nifedipine), 145 (felodipine); with PVP 127, 201 kJ/molreproduces ≈122 kJ/mol from plot readings marsac2006a E2, R3
wanapun2010 (outside)activation energy of griseofulvin crystallization by SHG140 ± 20 kJ/molreproduces [wanapun2010 E5]
tian2017melt viscosity of indomethacin with 10% HPMC, HPMCAS, PVP, 80–130 °CEη 159.7, 143.5, 190.5 kJ/molreproduces 159.7 kJ/mol from the printed slope tian2017 E1, R13, R14

Where it fails or is disputed

The Arrhenius plot of the 2006 paper has no rate unit and temperatures only on the 1/T axis, so it serves for slopes only marsac2006a X2. The activation-energy difference between the two drugs is significant only below Tg marsac2006a X6. The viscosity of pure indomethacin could not be measured because it crystallized, so the polymer melts have no reference tian2017 X12.

Concepts

kinetic barrier to nucleation, nucleation rate, accelerated stability testing. Findings. F2, F4, F29.

← Concepts and formulas

formula · as used in Crystallization tendency

Barandiaran-Colmenero critical cooling rate

Expression

\ln(\text{rate}) = A - \frac{B}{(\Delta T)^2},\quad \Delta T = T_m - T_{crys,max} baird2010 E1;

R_{crit} = \exp(A) baird2010 E2; baird2013 E1, with A the intercept of \ln(\text{cooling rate}) regressed on 1/(\Delta T)^2 baird2013 E3.

Variables and units

symbolmeaningunits
\text{rate}cooling rateK min⁻¹ baird2010 E1
\Delta Tundercooling at the crystallization peak on cooling, T_m - T_{crys,max}K baird2010 E1; baird2013 E4
A, Bintercept and slope of the regressionB in K² baird2010 E1
R_{crit}critical cooling rate, ΔT extrapolated to infinityK min⁻¹ baird2010 E2; baird2013 E1

Assumptions and valid domain

Undercooling must rise by an experimentally significant amount with cooling rate baird2010 E1; a higher cooling rate must lower the crystallization temperature baird2013 C17. Fitted over 5–20 °C/min baird2010 R13 and over about 5 to above 500 °C/min baird2013 X16.

Use in the library

paperused forvalues or fitted parametersstatus / recomputed
baird2010Rcrit of 19 class I moleculesr² 0.74–0.99 for 7 and below 0.55 for 12; Rcrit from 20 K/min (haloperidol) upwardreproduces (carbamazepine B 701 against 708, A 4.75 against 4.8; haloperidol \exp(3.0) = 20.1) baird2010 R13, E1, E2
baird2013Rcrit,cool of class I-A from 5–800 °C/min2.7 × 10³ (phenacetin) to 8.1 × 10³ K/min (4-biphenylmethanol), r² 0.89–0.99reproduces for phenacetin; haloperidol 6.6 × 10³ does not follow from A = 6.5 (6.65 × 10²); felbinac not reproducible baird2013 R7, X2, X13

Where it fails or is disputed

Over 5–20 °C/min the estimates contradicted experiment: haloperidol and phenacetin, with the lowest estimates, crystallized on liquid-nitrogen quench, whereas carbamazepine and griseofulvin, with higher estimates, quenched amorphous baird2010 C21; the approach "may be of limited utility" for organic molecules baird2010 C20, and accurate estimates need rates far below Rcrit and spanning several orders baird2010 R31, C22. A wider rate range seems to improve fits baird2013 C20, but the method is clearly inappropriate where undercooling rises and falls with rate (benzamide, lidocaine, indoprofen) baird2013 C16. Knowing that Rcrit,cool exceeds 500 °C/min is "probably sufficient" for pharmaceutical use baird2013 C21.

Concepts

Barandiaran-Colmenero method, critical cooling rate, degree of supercooling, glass-forming ability. Findings. F11.

← Concepts and formulas

formula · as used in Crystallization tendency

Burger-Ramberger heat-of-fusion rule

Expression

Not printed in either paper: applied in words from the relative melting points and enthalpies of fusion trasi2014b E2; named as the heat-of-fusion rule lu2016 E3.

Variables and units

symbolmeaningunits
T_m of each formmelting points of the two polymorphs°C trasi2014b M1, R6; lu2016 R20
\Delta H_{fus} of each formenthalpies of fusion of the two polymorphsJ/g trasi2014b R6; kJ/mol lu2016 R20

Assumptions and valid domain

None stated beyond the citations (refs 19 and 28) trasi2014b E2; lu2016 E3. The rule relates the order of melting points and enthalpies of fusion to monotropy or enantiotropy lu2016 E3.

Use in the library

paperused forvalues or fitted parametersstatus / recomputed
trasi2014bnilutamide stable form against form Istable Tm about 153 °C, about 98 J/g; form I 141 °C, about 66 J/g; monotropicnamed only trasi2014b E2, M1, R6, C11
lu2016vemurafenib α against βα 271 °C, 53.8 kJ/mol; β 248 °C, 33.3 kJ/mol; monotropicnamed only lu2016 E3, R20, C50

Where it fails or is disputed

The vemurafenib α enthalpy differs between places in the paper (51.0, 53.8, 55.9) lu2016 X1, though the order of the two forms does not change. For vemurafenib the Hoffman curves give a virtual transition at 319 °C, above both melting points, which supports monotropy lu2016 C51. The nilutamide free energy difference of about 3.4 kJ/mol rests on a relation that is not printed trasi2014b X13.

Concepts

Burger-Ramberger rules, monotropy, polymorphism. Findings. F7.

← Concepts and formulas

formula · as used in Crystallization tendency

Cabral critical cooling rate

Expression

R_{crit} = \exp\left(A^* - \frac{B^*}{T_m^2}\right) baird2010 E3;

R_{crit} = \exp\left[A^* - \frac{B^*}{T_m^2}\right] baird2013 E2.

Variables and units

symbolmeaningunits
A^*, B^*the intercept and slope of the Barandiaran–Colmenero regressionB* in K² baird2010 E3; baird2013 E2
T_mmelting pointK baird2010 E3; baird2013 E2
R_{crit}critical cooling rateK min⁻¹ baird2013 E1

Assumptions and valid domain

The undercooling is restricted to 0 to Tm − Tg instead of being extrapolated to infinity baird2010 E3; otherwise as for the Barandiaran–Colmenero form baird2013 E2.

Use in the library

paperused forvalues or fitted parametersstatus / recomputed
baird2010second Rcrit estimate for class Igriseofulvin 579 K/min; up to 3.26 × 10⁴ K/min (4-biphenylmethanol)reproduces: 588 \times \exp(-3711/491.15^2) = 579 baird2010 E3, R13
baird2013Rcrit,cool of class I-Aphenacetin 2.7 × 10³ K/minreproduces (2.68 × 10³); haloperidol printed 6.5 × 10³ against 6.47 × 10² recomputed baird2013 E2, R7, X2

Where it fails or is disputed

For griseofulvin B^*/T_m^2 = 3711/241228 = 0.0154, so the Cabral value is about 1.5% below the Barandiaran–Colmenero value (synthesis, from baird2010 E3); the two estimates are nearly equal (carbamazepine 118 and 117 K/min baird2010 R13) and share the same failures baird2010 C20, C21; baird2013 C16.

Concepts

critical cooling rate, Barandiaran-Colmenero method. Findings. F11.

← Concepts and formulas

formula · as used in Crystallization tendency

classical nucleation barrier

Expression

W^* = \frac{16\pi\gamma^3}{3\,\Delta G_v^2} baird2010 E24; vaneerdenbrugh2010a E3;

W^* = \frac{16\pi\,\sigma_{cm}^3}{3\,\Delta G_v^2} baird2012 E2, which enters the steady-state rate I_{SS} = I_0 \exp\left[-\frac{W^* + \Delta G_\eta}{k_B T}\right] baird2012 E1.

\Delta G^{*} \propto \frac{1}{(\ln S)^{2}} hsieh2012 E8. The same dependence appears in the nucleation-rate expression used for celecoxib, J = A\exp[-16\pi\gamma^{3}v^{2}/(3k^{3}T^{3}(\ln S)^{2})] xie2015 E3.

Variables and units

symbolmeaningunits
W^*free energy to form a nucleus of critical sizenot stated baird2010 E24
\gamma, \sigma_{cm}interfacial free energy of the nucleus–liquid (or nucleus–matrix) interface per areanot stated vaneerdenbrugh2010a E3; baird2012 E2
\Delta G_vfree energy difference per volume between crystal and liquidJ/cm³ vaneerdenbrugh2010a E2, E5
\Delta G^{*}free energy of forming the critical nucleusJ hsieh2012 E8
Ssupersaturation ratiodimensionless
\gamma, vinterfacial tension and molecular volume (in the full expression)J/m²; m³ xie2015 E3

Assumptions and valid domain

Homogeneous nucleation and a similar interfacial energy for the compared classes baird2010 E24; spherical nuclei baird2012 E2. Neither γ nor σcm was measured or estimated in any of the three papers baird2010 E24; vaneerdenbrugh2010a E3; baird2012 E2. In the supersaturation form the barrier falls steeply as S rises, so crystals nucleate only above a certain supersaturation, which depends on compound and conditions hsieh2012 C3; no numerical output is stated in the pH-shift paper.

Use in the library

paperused forvalues or fitted parametersstatus / recomputed
baird2010argument that a more negative ΔGv of class I lowers the barriernoneinputs not printed baird2010 E24, C61
vaneerdenbrugh2010areading the ΔGv–class linknoneinputs not printed vaneerdenbrugh2010a E3, C39
baird2012argument that viscosity enters nucleation and growthnoneinputs not printed baird2012 E2, C4, C32
duong2022bposaconazole barrier in buffer and nucleus size from fitted γ\Delta G^{*} = 16\pi\gamma^{3}\nu^{2}/(3k^{2}T^{2}\ln^{2}S); 2.73 × 10⁻²¹ J against ≈2.72 × 10⁻²¹ J read from the figure; PBS 2.72 × 10⁻²¹ J at S 33.6reproduces duong2022b E8, R17
hsieh2012why type 1 bases precipitate only after reaching a supersaturation of 1.4–26.7 times the titration solubilitynonenot applicable (no numerical output) hsieh2012 E8, R5
xie2015reading polymer effects on celecoxib induction times (ASD dissolution chapter)γ, v not fittedprinted xie2015 E3

Where it fails or is disputed

W* depends on γ³ but only on ΔGv²; the authors read the ΔGv–class correlation as suggesting that γ differences between compounds are small vaneerdenbrugh2010a C39, an inference that rests on a ΔGv ranking that is not monotonic across classes vaneerdenbrugh2010a X10. In the pans and films used, nucleation may be heterogeneous baird2010 C19; baird2012 C8, which the homogeneous form does not cover. Precipitation was seen at widely different supersaturations (1.4 for pyrimethamine to 26.7 for droperidol) hsieh2012 R5, C22. (synthesis: the barrier expression alone does not predict this spread without the interfacial energies of the compounds, which the paper does not give.)

Concepts

classical nucleation theory, nucleation, thermodynamic driving force for crystallization, heterogeneous nucleation. Findings. Crystallization tendency: F3, F6. Crystallization from solution: F1, F14, F34.

← Concepts and formulas

formula · as used in Crystallization tendency

classical nucleation theory

Expression

J \propto \exp\left(-\frac{\Delta G^{*}}{k_B T}\right) hsieh2012 E7

J = A\exp\left(\frac{-\Delta G_{crit}}{kT}\right) ilevbare2013c E4

J_{N} = v^{*} z n \exp\left(\frac{-16\pi\gamma^{3}v^{2}}{3k_{B}^{3}T^{3}(\ln S)^{2}}\right) [indulkar2018 E6]

J = A\exp\left(\frac{16\pi \nu_o^{2} \gamma^{3}}{3 (k_B T)^{3} (\ln S)^{2}}\right) [elkhabaz2021 E2] (printed without the minus sign)

J = A\,S\,\exp\left(\frac{-16\pi\gamma^{3}\nu^{2}}{3k^{3}T^{3}\ln^{2}S}\right) duong2022b E6

\ln(J/S) = \ln A - \frac{B}{\ln^{2}S}, \qquad B = \frac{16\pi\gamma^{3}\nu^{2}}{3k^{3}T^{3}} duong2022b E7

J = A\exp\left(-\frac{16\pi\gamma^{3}v^{2}}{3k^{3}T^{3}(\ln S)^{2}}\right) trasi2015 E6; xie2015 E3

Nucleation rate with a thermodynamic and a kinetic barrier, as printed in the solid-state papers of the physical-stability chapter:

I = A\exp\left(\frac{-\left(\Delta G^{*} + \Delta G_{a}\right)}{kT}\right) konno2006 E8

I = A\exp\frac{-\left(\Delta G^{*} + \Delta G_{a}\right)}{kT} marsac2006a E9

and for the undercooled melt:

I_{SS} = I_0\exp\left(-\frac{W^{*} + \Delta G_{\eta}}{k_B T}\right) baird2012 E1

In the reduced-temperature form printed for acetaminophen, with its typesetting as printed:

J = A\exp\left[-\frac{16\pi r^{3}}{3kT_m\Delta H_f^{2}T_r(\Delta T_r)^{2}} + \frac{\Delta G'}{KT}\right] trasi2012a E1

where the card reads r as the interfacial energy \gamma and the kinetic term as -\Delta G'/kT, as the text describes trasi2012a X1.

The ASD dissolution papers of 2008 and 2010 print it with ln S unsquared,

J = A\exp\left(-\frac{16\pi\gamma^{3}v^{2}}{3k^{3}T^{3}\,(\ln S)}\right) [konno2008a E3],

J \propto \exp\left[-\frac{1}{\ln S}\right] alonzo2010 E4;

the ritonavir dispersion paper states the dependence in words only: the relative nucleation rate rises with the drug concentration in a phase and falls with the crystal–solution interfacial tension [ilevbare2013d E1].

Variables and units

symbolmeaningunit
Jnucleation ratem⁻³ s⁻¹
Apre-exponential kinetic factor, depends on molecular mobilitym⁻³ s⁻¹
ΔG*, ΔG_critfree energy of forming a critical nucleusJ
k, k_BBoltzmann constantJ/K
Tabsolute temperatureK
γinterfacial tension (free energy) between nucleus and solutionJ/m² (mJ/m² in the posaconazole fits)
ν, v, ν_omolecular volumem³ (920 ų for posaconazole)
Ssupersaturationdimensionless
v*, z, nmonomer attachment rate, Zeldovich factor, number density of moleculesas printed
Inucleation rate in the film papersm⁻³ s⁻¹ konno2006 R4
\Delta G^{*}, W^{*}thermodynamic barrier, from interfacial term against bulk driving forceJ konno2006 E8
\Delta G_{a}, \Delta G_{\eta}kinetic barrier for transport onto the nucleusJ konno2006 E8
T_r, \Delta T_rreduced temperature T/T_m and 1 - T_r– trasi2012a E1
\Delta H_f; \Delta G'heat of fusion; activation energy for molecular motionnot given trasi2012a E1

Assumptions and valid domain

Clusters grow by sequential molecular additions; the barrier rises with the cube of the interfacial energy and falls with (\ln S)^{2} (see classical nucleation barrier) ilevbare2013c E4, C14; hsieh2012 E8. Used qualitatively; no fit to induction times is printed. The solid-state papers assume homogeneous nucleation, although nucleation in the films is "most likely heterogeneous"; stresses from the volume change are ignored konno2006 C33; marsac2006a C32. In the ASD dissolution papers of 2008–2013 it is used only qualitatively: higher supersaturation, faster nucleation [konno2008a C20; alonzo2010 C43; ilevbare2013d C40]. The 2015 celecoxib paper prints the classical form with (\ln S)^{2} and uses it to argue that neither supersaturation nor interfacial tension explains the polymer differences in induction time xie2015 E3, C43.

Use in the library

paperused forvalues or fitted parametersstatus / recomputed
hsieh2012why type 1 bases precipitate only above a certain supersaturationnonenot applicable (no numerical output) hsieh2012 E7
ilevbare2013cwhy a polymer gains more at lower supersaturation (celecoxib 31-fold at S 7.3)noneinputs not printed ilevbare2013c E4, C45
xie2015celecoxib dispersions: J = A\exp[-16\pi\gamma^{3}v^{2}/(3k^{3}T^{3}(\ln S)^{2})]; longer induction times with polymers read as lower nucleation rate; why HPMCAS and HPMC lengthen celecoxib induction times more than PVPγ, v not fitted; γ, v, A not printedprinted; inputs not printed xie2015 E3, C42, C43
trasi2015J = A exp[−16πγ³v²/(3k³T³(ln S)²)] to state that T, S and γ set Jacetaminophen S 1.5, 20 °Cinputs (A, γ, v) not printed trasi2015 E6
li2016anamed only: nucleation rate depends highly on supersaturationbile-salt seriesnamed only [li2016a E5]
indulkar2018why lecithin might accelerate nucleation through γnoneinputs not printed [indulkar2018 E6, C40]
elkhabaz2021why surfactants should lower γ, and why human fluid does not fitnoneprinted without minus sign [elkhabaz2021 E2, X1]
duong2022blinearized fit of posaconazole induction times in buffer and three mediaγ 3.52, 2.61, 2.73, 3.24 mJ/m²; A 2.44, 5.14, 4.98, 9.87reproduces from the printed slopes and intercepts duong2022b E7, R16
konno2006sorting polymer effects into driving force and kinetic barriernoneinputs not printed konno2006 E8
marsac2006aframework for the nifedipine–felodipine comparisonnoneframework only marsac2006a E9
baird2012role of viscosity in class differencesnoneinputs not printed baird2012 E1
trasi2012aframework for polymer effects on acetaminophen nucleationnoneinputs not printed; printed form has misprints trasi2012a E1, X1
konno2008afelodipine nucleates readily at a high solubility rationonenot applicable [konno2008a E3]
alonzo2010scheme of dissolution against crystallizationnonenot applicable alonzo2010 E4
ilevbare2013dnucleation favoured in drug-rich dropletsnoneprinted in words [ilevbare2013d E1]

Where it fails or is disputed

Induction times include growth to detectable size, so a rate from them is nucleation plus early growth ilevbare2013c X14. The two-step picture (disordered cluster, then reorganization) is assumed to be rate-limiting but not shown ilevbare2013c C5, C15. The critical supersaturation at which precipitation is seen depends on compound and conditions hsieh2012 C3. The Poisson fit uses the printed volume 0.0012 m³ and a positive exponent, both misprints (V = 1.2 mL and a minus sign reproduce the 95% times within about 12%) trasi2015 X1, X2, E5. In the posaconazole fits the high-supersaturation sets hold 3–9 points (R² 0.84–0.97, FeSSIF V2 not fittable), and the conversion of induction time into J is not stated duong2022b X10, X11. No parameter was fitted in the solid-state papers; the theory serves to exclude a driving-force route and to frame a kinetic one konno2006 C35, C40. The polymer effects on acetaminophen nucleation were "not particularly straightforward" to explain with it, and it does not contain molecular recognition trasi2012a C49. The 2008 and 2010 dissolution papers print ln S without the square of the classical expression; use the squared form to compute rates [konno2008a X8; alonzo2010 X3]. The 2015 celecoxib form carries the square; in its argument the supersaturation is not the same with and without polymer (about 14.7 against 10.3–11.6), and the text refers to Eq. 2 where interfacial tension appears in Eq. 3 xie2015 X9, X1.

Concepts

classical nucleation theory, nucleation rate, crystal-solution interfacial free energy, kinetic barrier to nucleation, interfacial free energy, nucleation, interfacial tension. Findings. Crystallization from solution: F1, F14, F27, F29, F5, F34, F39, F42. Physical stability: F1, F4, F17. Supersaturation from ASD dissolution: F1, F4, F5, F13.

← Concepts and formulas

formula · as used in Crystallization tendency

critical cooling rate to avoid nucleation

Expression

R_{Ncrit} \sim \frac{1}{\tau_{Nmin}} baird2010 E21, with the nucleation time \tau_N = \frac{1}{J_N V} baird2010 E22.

Variables and units

symbolmeaningunits
R_{Ncrit}critical cooling rate to avoid nucleationnot stated baird2010 E21
\tau_{Nmin}minimum nucleation time between Tm and Tgnot stated baird2010 E21
J_N, Vnucleation rate and sample volumenot stated baird2010 E22

Assumptions and valid domain

An approximation baird2010 E21. Glass formation requires cooling faster than the maximum crystallization rate; the minimum nucleation time lies between Tm and Tg, where the driving force rising with undercooling meets the viscosity that restricts diffusion near Tg baird2010 C49.

Use in the library

paperused forvalues or fitted parametersstatus / recomputed
baird2010argument for why nucleation and growth windows decide glass formationnoneinputs not printed baird2010 E21, E22, C49

Where it fails or is disputed

No nucleation time or rate was measured baird2010 E21. Because τN depends on sample volume baird2010 E22, the critical rate is not a pure material property (synthesis: from the 1/(J_N V) form).

Concepts

critical cooling rate, nucleation, nucleation temperature zone, glass-forming ability. Findings. F3, F16.

← Concepts and formulas

formula · as used in Crystallization tendency

crystal growth inhibition effectiveness ratio

Expression

The polymer effectiveness ratio for crystal growth inhibition is the growth rate without polymer over the rate with polymer:

E_G=\frac{R_{G,np}}{R_{G,p}}

[moseson2020a E2; moseson2021 E1]. E_G>1 means the polymer slows growth.

The crystallization-from-solution papers print the same ratio as

E_g = \frac{R_{g0}}{R_{gp}}

[ilevbare2012c E5; ilevbare2012a E3; ilevbare2012b E12; ilevbare2013b E5]. The inverse form R/R_0 = R_{gp}/R_{g0} is used in the Kubota–Mullin model [ilevbare2012b E7].

For the melt growth of two felodipine polymorphs it is printed in words: the bulk growth rate without polymer divided by the bulk rate with 3% PVP, per polymorph and temperature kestur2013a E2.

Variables and units

symbolmeaningunitids
E_Geffectiveness ratiodimensionless[moseson2020a E2]
R_{G,np}crystal growth rate without polymerµg/s (vessel mass rate)[moseson2020a E2, X12]
R_{G,p}crystal growth rate with polymerµg/s[moseson2020a E2]
R_{g0}initial growth rate without polymerµg mL⁻¹ min⁻¹ or µg min⁻¹[ilevbare2012a E3]
R_{gp}initial growth rate with polymer or additivesame[ilevbare2012a E3]
E_geffectiveness ratio; 3 means growth three times slowerdimensionlessilevbare2013b E5
bulk growth rates (felodipine melt)linear growth rates without and with 3% PVP, read from plotsm/skestur2013a E2, R8, R13

Assumptions and valid domain

Evaluated at the highest seed level (50 µg/mL) at S = 6 for indomethacin; for bicalutamide from seeded growth at 150 µg/mL [moseson2020a E2, P20; moseson2021 R5]. The growth rates are vessel mass rates not normalised to seed surface area and the extraction method is not printed [moseson2020a X12]. In the seeded-growth papers a ratio above 1 is taken as effective and below 1 as accelerating; the rate is the slope of the dissolved drug over the first 2 min at fixed seed mass and supersaturation [ilevbare2013b C17; ilevbare2012a E3, E4]. Tested for ritonavir, celecoxib and efavirenz at 37 °C, pH 6.8, 5 µg/mL polymer. In the felodipine polymorph work it is applied to growth from the supercooled melt at 60–100 °C, not from solution kestur2013a R15, C28.

Use in the library

paperused forvalues or fitted parametersstatus / recomputed
moseson2020aindomethacin with 5 and 50 µg/mL PVPVAE_G 18 and 56reproduces from the figure readings: 0.99 / 0.055 = 18 and 0.99 / 0.018 = 55 [moseson2020a R5, E2]
moseson2021bicalutamide with PVPVA 70 µg/mL and HPMCAS 1 mg/mLE_G 8.6 and 158inputs not printed (growth rates are in a supplement that was not supplied) [moseson2021 R5, E1]
ilevbare2012c34 polymers on ritonavir0.83–3.25absolute rates not printed [ilevbare2012c E5, R9]
ilevbare2012bpolymers against S and pH6.40, 2.99, 1.56; 2.98 → 1.12not applicable (definition) [ilevbare2012b E12]
ilevbare2012a14 polymers, 13 pairs, surfactantsup to 12.1; 0.49 for TPGSreproduces (3.6/1.1 = 3.27, printed 3.3) [ilevbare2012a E3]
ilevbare2013b16 polymers on three drugsup to 12.6reproduces (69.80/11.88 = 5.88, text about 6.0) ilevbare2013b E5
schram2015E_g = R_0/R_P of felodipine with HPMCAS-LF2.29 (pH 6.8), 1.28 (pH 3)slopes not tabulated; end-point estimate gives 2.3 and 1.3 [schram2015 E4, R4]
schram2016same, R_0 of buffer-grown seeds without polymer2.7, 5.5, 8.0, 12.0 with HPMCAS in solution; 1.8, 3.0, 8.4 withoutplot readings [schram2016 E5, R2]
lu2017aE_G at δ 2.4 and E_C at δ 7.9 or 12.5 for telaprevir with bile saltsE_G 1.0–4.6; STCDC no desupersaturationplot readings [lu2017a E2, E3, R7, R8]
lu2018telaprevir seeds with bile salt mixtures, δ 7.9E_G = R_0/R over the first 10 minSTDC 3.8; STCDC >60; binary-L 35.5; binary-H 7.4; inputs not printed [lu2018 E3, R5]
kestur2013awhether PVP acts in the liquid or at the crystal faces of felodipine forms I and IIform I ≈15.8 → ≈3.5, form II ≈18.7 → ≈6.5 (60 → 100 °C)reproduces from Figs. 2 and 3 for form II and for form I at 90–100 °C, not for form I at 60–80 °C kestur2013a E2, X7
kestur2013bsurface against bulk growth of felodipine form I with 3% PVP≈2 (surface), ≈7 (bulk)from the printed rates kestur2013b R12

Where it fails or is disputed

The ratio falls as supersaturation rises, depends on how the seeds were prepared and on the polymer concentration, so values from different systems are not comparable [ilevbare2012b C56; schram2016 C13]. A large ratio in solution does not predict protection against crystallisation in the solid [moseson2021 C51]. The stated rule (E_g above 1 is effective) does not match the counts of effective polymers or the polymers called ineffective at 1.0–1.4 (cut-off near 1.2 in practice) [ilevbare2013b X3; ilevbare2012c X7]. Rates depend on seed mass, volume and methanol fraction, which are not always printed [ilevbare2012c X16]. A polymer that raises the solubility lowers S and flatters the ratio (F19) ilevbare2013c X7. The reference rate R_0 differs: one seed route in the felodipine seed study and the pH-independent single curve in the 2015 paper [schram2016 E5; schram2015 X3]; rates are not normalised to area [schram2016 X12]. In the felodipine melt work the error bars of the two polymorphs separate at 90–100 °C, so equality of their ratios holds within about a factor of two kestur2013a X6.

Concepts

crystal growth inhibition, crystallization inhibition by polymers, crystal seeding, crystal growth rate, polymorphism. Findings. ASD process: F25, F27, F29. Crystallization from solution: F7, F8, F9, F17, F18, F23, F24, F33. Physical stability: F3, F19.

← Concepts and formulas

formula · as used in Crystallization tendency

depth of field of a microscope objective

Expression

As printed in the four nucleation papers:

D_{tot} = \frac{\lambda n}{NA^{2}} + \frac{n e}{M \cdot NA} konno2006 E1; marsac2006a E1; konno2008 E1; marsac2008a E2

Variables and units

symbolmeaningunits
D_{tot}total depth of field, used as the sampled thicknessmm (0.0144 mm at 10×) konno2006 E1
\lambdawavelength of light550 nm konno2006 E1
nrefractive index of the medium1.000 (air) konno2006 E1
NAnumerical aperturenot printed konno2006 E1
esmallest distance resolved by the detector14 µm konno2006 E1
Mlateral magnification10× konno2006 E1

Assumptions and valid domain

The counted nucleation sites lie within the depth of field, so the site density per volume follows from the density per area and D_{tot} konno2008 C38. Used for birefringent sites of about 1 µm or more in spin-coated films konno2008 C38.

Use in the library

paperused forvalues or fitted parametersstatus / recomputed
konno2006volume nucleation rates of felodipine films0.0144 mm at 10×NA not printed; NA = 0.25 reproduces 14.4 µm konno2006 E1
marsac2006anifedipine and felodipine nucleation rates0.0144 mm at 10×same check marsac2006a E1
konno2008nucleation at 0–75% RH0.0144 mmsame check konno2008 E1
marsac2008anucleation with PVP and water0.0144 mm at 10×; other objectives not givensame check marsac2008a E2, X10

Where it fails or is disputed

The 2006 and 2008 felodipine papers describe the conversion as multiplication by D_{tot} where a per-volume density needs division konno2006 X6; konno2008 C38. Rates are therefore comparable only within this counting method; the film thickness is not printed marsac2006a X10; konno2008 X7.

Concepts

nucleation rate, nucleation. Findings. F1.

← Concepts and formulas

formula · as used in Crystallization tendency

entropy of fusion

Expression

\Delta S_{fus} = \frac{\Delta H_{fus}}{T_{melt,ons}} baird2010 E26;

\Delta S_{fus} = \frac{\Delta H_{fus}}{T_m} vaneerdenbrugh2010a E1.

Variables and units

symbolmeaningunits
\Delta S_{fus}entropy of fusionkJ mol⁻¹ K⁻¹ baird2010 E26; J cm⁻³ K⁻¹ after volume conversion vaneerdenbrugh2010a E1
\Delta H_{fus}heat of fusion from DSCkJ mol⁻¹ baird2010 E26; J cm⁻³ vaneerdenbrugh2010a E1
T_m, T_{melt,ons}melting point (onset)K baird2010 E26; vaneerdenbrugh2010a E1

Assumptions and valid domain

None stated beyond DSC inputs baird2010 E26; vaneerdenbrugh2010a E1. The 31-drug growth study uses the melting entropy ΔSm per mole in \Delta G \approx \Delta S_m\,\Delta T, an approximation trasi2014a E3.

Use in the library

paperused forvalues or fitted parametersstatus / recomputed
baird2010descriptor tables and class PCAhaloperidol 12.8 × 10⁻² kJ mol⁻¹ K⁻¹; class I contribution +0.41reproduces: 54.26/425.15 = 0.1276 baird2010 E26, R22
vaneerdenbrugh2010aclass PCAcontributions +0.46, −0.27, −0.33 for classes I, II, IIIinputs not printed vaneerdenbrugh2010a E1, R20
marsac2006afelodipine and nifedipine74.3 and 89.6 J mol⁻¹ K⁻¹reproduces marsac2006a E11

Where it fails or is disputed

On a volume basis the fast class I has the higher entropy of fusion baird2010 R22; vaneerdenbrugh2010a R20, while on a molar basis the fastest growers had lower melting entropy and the normalized growth rate fell with ΔSm/R trasi2014a R8, R11. (synthesis: the two bases and the two outcomes, class and growth rate, differ, so the signs need not conflict; class I molecules are small, which raises per-volume values.)

Concepts

entropy of fusion, enthalpy of fusion, thermodynamic driving force for crystallization. Findings. F6, F13.

← Concepts and formulas

formula · as used in Crystallization tendency

Flory-Huggins activity equation

Expression

Activity coefficient of the drug in a polymer, with the amorphous drug as the solvent (2006):

\ln\gamma_{\mathrm{drug}} = \ln\frac{\Phi_{\mathrm{drug}}}{x_{\mathrm{drug}}} + \left(1-\frac{1}{m}\right)\Phi_{\mathrm{polymer}} + \chi\,\Phi_{\mathrm{polymer}}^{2} marsac2006b E6;

the same quantity scaled from a liquid analogue (1-ethyl-2-pyrrolidone, EP) to PVP (2009):

\ln\gamma_{\mathrm{drug}}^{PVP} = \frac{MV_{\mathrm{drug}}}{MV_{\mathrm{lattice}}}\left[\frac{1}{m_{\mathrm{drug}}}\ln\frac{\Phi_{\mathrm{drug}}}{x_{\mathrm{drug}}} + \left(\frac{1}{m_{\mathrm{drug}}}-\frac{1}{m_{PVP}}\right)\Phi_{PVP}\right] + \ln\gamma_{\mathrm{drug}}^{EP} marsac2009 E8;

\ln\gamma_{\mathrm{drug}}^{PVP} = \frac{MV_{\mathrm{drug}}}{MV_{\mathrm{lattice}}}\left[\frac{1}{m_{\mathrm{drug}}}\ln\frac{\Phi_{\mathrm{drug}}}{x_{\mathrm{drug}}} + \left(\frac{1}{m_{\mathrm{drug}}}-\frac{1}{m_{PVP}}\right)\Phi_{PVP} + \chi\,\Phi_{PVP}^{2}\right] marsac2009 E9.

As the activity of the drug in the library's other papers:

\ln a = \ln\Phi_1 + \left(1-\frac{1}{x_{12}}\right)\Phi_2 + \chi\,\Phi_2^{2} konno2006 E9;

\ln a_{\mathrm{drug}} = \ln\Phi_{\mathrm{drug}} + \left(1-\frac{1}{m}\right)\Phi_{PVP} + \chi\,\Phi_{PVP}^{2} marsac2006a E10.

The ASD dissolution papers print it as the change of the drug chemical potential in an undissolved polymer:

\frac{\Delta\mu_{\mathrm{drug}}}{RT} = \ln\Phi_{\mathrm{drug}} + \left(1-\frac{1}{r}\right)\Phi_{\mathrm{poly}} + \chi\,\Phi_{\mathrm{poly}}^{2} [li2018 E4]

Variables and units

symbolmeaningunits
\gamma_{drug}, aactivity coefficient and activity of the drugdimensionless marsac2006b E6; konno2006 E9
\Phivolume fractiondimensionless marsac2006b E6
x_{drug}mole fraction of the drugdimensionless; mole-fraction basis in PVP not printed marsac2009 X13
mratio of polymer to lattice-site volumedimensionless marsac2006b E6
MVmolar volumecm³/mol (e.g. 288.62 for nifedipine) marsac2006b M1; marsac2009 E8
\chiinteraction parameterdimensionless marsac2006b E6
\Delta\mu_{\mathrm{drug}}change of the drug chemical potential on mixingJ/mol [li2018 E4]
\Phi_{\mathrm{drug}}, \Phi_{\mathrm{poly}}volume fractions of drug and polymer (2018 notation)dimensionless [li2018 E4]
rV_{\mathrm{poly}}/V_{\mathrm{drug}}, volume ratio of polymer to drug (2018 notation for m)dimensionless [li2018 E4]

Assumptions and valid domain

In the analogue liquid the first two terms vanish when the mixing entropy is ideal (minimal size difference) marsac2006b C46. The 2009 scaling assumes ideal mixing entropy with EP, identical contacts of the drug with EP and with the PVP repeat unit, Flory–Huggins combinatorial entropy for PVP, and equal γ at the solubility limits in EP and in PVP marsac2009 E6, E7, C11. In the ASD dissolution use: a drug–polymer dispersion that is one phase and stays one phase on hydration, with a polymer largely insoluble in the medium; the change of χ with water is not included [li2018 C45, C46].

Use in the library

paperused forvalues or fitted parametersstatus / recomputed
marsac2006bsolubility of the crystalline drug in PVP and methyl pyrrolidonewith χ −3.8 and −4.2reproduces for felodipine in PVP K12 with m = 7.50 (Φ_drug 0.210, 0.31 g/g against 0.31); felodipine in K29/32 gives 0.29 against 0.25 marsac2006b E6, R17
marsac2009γ of five compounds in PVP K12, K29/32, K90 from γ in EPγ in PVP 0.50–5.97inputs not printed (lattice volume of EP, mole-fraction basis) marsac2009 E8, E9, R9
marsac2006adrug activity against PVP content with χ of the melting routeactivity ≈0.84 (nifedipine) and ≈0.83 (felodipine) at Φ_PVP 0.2reproduces approximately (0.839, 0.826 with 1/m ≈ 0) marsac2006a E10, R19
marsac2006aargument that PVP lowers the driving force alike and little for nifedipine and felodipine, so the ranking persistsactivity ≈0.96, 0.84, 0.67 at Φ 0.1, 0.2, 0.3same calculation as the row above marsac2006a R19, C47
konno2006argument that Flory–Huggins gives little activity loweringnoneχ and x₁₂ for felodipine–polymer not given konno2006 E9, C35
li2018qualitative ranking of the suppression of lopinavir concentration by insoluble polymersχ and r not determinedinputs not printed [li2018 E4, C44, C51]

Where it fails or is disputed

The values in PVP are predictions under the stated assumptions; none was checked against a solubility measured in a polymer, and the one check (methyl pyrrolidone) is about 30% low for nifedipine marsac2006b X10; marsac2009 X17. The 2009 relation has no printed lattice volume, so it cannot be recomputed from the paper marsac2009 X4, X13. The combinatorial and residual parts are linked, because order imposed by exothermic interactions lowers the entropy of mixing marsac2009 C10. The conclusion drawn from it in the physical-stability papers, that a few percent of polymer acts kinetically, was not tested by any measured activity konno2006 C35; marsac2006a C46. For lopinavir, χ was not measured for any lopinavir–polymer pair, so the use is qualitative [li2018 C46, C51]. With χ = 0 and a large r the equation gives a drug activity of about 0.82 at a drug volume fraction of 0.5, above the proportional-dilution line that the paper calls ideal (extractor calculation) [li2018 X2].

Concepts

activity coefficient, combinatorial entropy of mixing, Flory-Huggins theory, drug solubility in polymer, thermodynamic driving force for crystallization, Flory-Huggins interaction parameter, chemical potential, solubility suppression. Findings. Miscibility and drug-polymer interactions: F8, F9. Physical stability: F4. Supersaturation from ASD dissolution: F18.

← Concepts and formulas

formula · as used in Crystallization tendency

Hoffman equation

Expression

\Delta G_v = -\frac{\Delta H_{fus}\,(T_{melt,ons} - T)\,T}{T_{melt,ons}^{2}} baird2010 E25;

\Delta G_v = -\frac{\Delta H_{fus}\,(T_{m} - T)\,T}{T_{m}^{2}} vaneerdenbrugh2010a E2;

\Delta G_v = \frac{\Delta H_{m}\,(T_{m} - T)\,T}{T_{m}^{2}} lu2016 E1.

Named but not printed in baird2012 E6. The sign differs between papers: the two 2010 papers print a leading minus sign, the 2016 paper does not and plots the negative of its own expression lu2016 X17.

Two more printed forms. In the PEG dispersions paper the expression is printed without a leading sign and used inside the simplified solubility equation \ln[\gamma_d x_d] = -\Delta G_{fus}/(R\,T) [baird2011 E5]:

\Delta G_{fus} = \frac{\Delta H_{fus,d}\,(T_{melt,d} - T)\,T}{T_{melt,d}^{2}} [baird2011 E6].

In the solution-methods paper it is printed with a positive sign and \Delta T = T_m - T:

\Delta G = \frac{\Delta H_f\,\Delta T}{T_m}\cdot\frac{T}{T_m} raina2015a E2,

which equals \Delta H_f\,(T_m - T)\,T/T_m^{2} and is the amorphous-minus-crystal free energy that enters e^{\Delta G/RT} of the amorphous solubility estimate.

The papers of the amorphous solubility chapter print the relation with a positive sign, as the amorphous-minus-crystal free energy that enters e^{\Delta G/RT}:

\Delta G_{a\to c} = \frac{\Delta H_f\,(T_m - T)\,T}{T_m^{2}} [ilevbare2013a E2; indulkar2015 E3; mosqueragiraldo2015 E2; almeidaesousa2016 E2];

the 2015 Almeida e Sousa paper writes \Delta_{fus}H for the heat of fusion and \Delta G_{c\to a} for the direction almeidaesousa2015 E2. For atazanavir it is printed again,

\Delta G = \frac{\Delta H_f\,(T_m - T)\,T}{T_m^{2}} [indulkar2017a E2].

The papers of the crystallization-from-solution chapter print it with a positive sign as well:

\Delta G_c = \Delta H_f\,\frac{(T_m - T)\,T}{T_m^{2}} hsieh2012 E3; hsieh2014 E2;

\Delta G_{a \to c} = \frac{\Delta H_{fus}\,(T_m - T)\,T}{T_m^{2}} [ilevbare2012b E9; ilevbare2013c E7];

\Delta G_C = \frac{\Delta H_f\,(T_m - T)\,T}{T_m^{2}} ilevbare2013b E3;

\Delta G = \frac{\Delta H_f\,\Delta T\,T}{T_m^{2}}, \qquad \Delta T = T_m - T [jackson2014 E2].

The moisture-induced separation paper of the physical-stability chapter prints it with a positive sign,

\Delta G_c = \Delta H_f\frac{\left(T_m - T\right)T}{T_m^{2}} [rumondor2009c E3],

and the felodipine kinetics paper names it without an expression [rumondor2009b E1].

The ASD dissolution papers print it as

\Delta G_c = \Delta H_f\,\frac{(T_m - T)\,T}{T_m^{2}} [konno2008a E2],

\Delta G = \frac{\Delta H_f\,\Delta T\,T}{T_m^{2}} [alonzo2010 E2; alonzo2011 E3],

and, with the sign that makes \Delta G negative,

\Delta G = -\frac{\Delta H_f\,(T_m - T)\,T}{T_m^{2}} duong2022a E2.

Variables and units

symbolmeaningunits
\Delta G_vfree energy difference between crystal and amorphous or liquid stateJ/cm³ after conversion with crystalline density vaneerdenbrugh2010a E2, E5; baird2010 E25; kJ/mol lu2016 E1
\Delta H_{fus}, \Delta H_menthalpy of fusionkJ/mol or J/cm³ baird2010 E26; vaneerdenbrugh2010a E1
T_m, T_{melt,ons}melting temperature (onset)K vaneerdenbrugh2010a E1; lu2016 E1
Ttemperature of evaluation293.15 K baird2010 E25; vaneerdenbrugh2010a E2, 298 K baird2012 E6, crystallization temperature lu2016 E1; 25 or 37 °C [hsieh2014 E2; ilevbare2012b E9]
\Delta G_{fus}, \Delta H_{fus,d}, T_{melt,d}free energy difference between crystalline and liquid drug at T (298.15 K in the check); heat of fusion of the drug in kJ/mol; its melting temperature\Delta H_{fus,d} in kJ/mol; temperatures in K; unit of \Delta G_{fus} not printed [baird2011 E5, E7]
\Delta H_fmolar heat of fusionnot printed raina2015a E2
\Delta TT_m - TK raina2015a E2
\Delta G_{a\to c}amorphous-minus-crystal free energy that enters e^{\Delta G/RT} in the amorphous solubility papers (\Delta G_{c\to a} in the 2015 Almeida e Sousa paper)kJ/mol in the use table below ilevbare2013a E2; almeidaesousa2015 E2
\Delta G_c, \Delta G_Cfree-energy difference between amorphous (supercooled liquid) and crystalkJ/mol hsieh2012 E3; ilevbare2013b E3
\Delta H_f, T_m, T in the moisture paperheat of fusion; melting and evaluation temperatureskJ/mol; K [rumondor2009c E3]
\Delta G_c, \Delta G (ASD dissolution papers)free-energy difference between amorphous and crystalline drugJ/mol [konno2008a E2; alonzo2010 E2]

Assumptions and valid domain

Lu et al. call it an approximation lu2016 E1. Papers outside this chapter state that it assumes enthalpies of crystal and supercooled liquid linear in temperature and applies above Tg marsac2006a E3; marsac2006b E4, that it assumes a constant heat-capacity difference and small undercooling almeidaesousa2015 E2, and that it may not suit large undercoolings mosqueragiraldo2015 E2. In this chapter it is evaluated at room temperature, far below Tm and below Tg for many compounds baird2010 E25; baird2012 E6. In the PEG dispersions paper it is used as an estimate of \Delta G_{fus} from thermal properties, with the drug solubility in PEG 400 as a proxy for the polymer [baird2011 E6, C20, C21]. In the solution-methods paper no assumptions are stated beyond the reference; ΔHf and Tm come from DSC at 10 °C/min raina2015a E2, P4. In the amorphous solubility papers the assumptions are a constant, temperature-independent ΔCp, small undercooling and Tg close to T∞ almeidaesousa2015 C8, and a temperature of interest "reasonably close to Tm", which is violated for telaprevir (Tm 238 °C, 37 °C) mosqueragiraldo2015 C24, M1. The pH-shift and crystal-growth papers state that the enthalpies of crystal and supercooled liquid vary linearly with temperature, "to a first approximation" [hsieh2014 E2; jackson2014 E2]; ΔHf and Tm come from DSC of the crystalline material and temperatures must be in kelvin hsieh2012 E3. The felodipine kinetics paper states that it applies to a pure amorphous drug; with polymer and sorbed water a reasonable estimate is "extremely difficult" [rumondor2009b C17]. In the ASD dissolution papers it is called increasingly approximate far below Tg (cited) [alonzo2011 C5] and was judged a good estimate for felodipine in earlier work (cited) [konno2008a R20].

Use in the library

paperused forvalues or fitted parametersstatus / recomputed
baird2010ΔGv descriptor in the class PCAclass I contribution −0.40 (more negative ΔGv)inputs not printed baird2010 E25, R22.g
vaneerdenbrugh2010aΔGv descriptor in the class PCAcontributions −0.29, +0.29, +0.13 for classes I, II, IIIinputs not printed vaneerdenbrugh2010a E2, R20.e
baird2012ΔGv column at 298 K−8.6 (lidocaine) to −54.7 J/cm³ (acetaminophen)named only; the standard form reproduces the other 33 rows within about 0.6 J/cm³, ritonavir printed −37.6 against about −21.4 baird2012 E6, M5, M12, X2
lu2016driving force of α and β vemurafenib; virtual transitionα 11.0 kJ/mol at 115 °C; transition 319 °Creproduces lu2016 E1, R22
marsac2006afelodipine and nifedipine driving force6.22 and 8.83 kJ/mol at 25 °Creproduces marsac2006a E3
ilevbare2013cdriving force at 37 °C (extractor calculation)ritonavir ≈10.2, efavirenz ≈2.70, celecoxib ≈7.74 kJ/molinputs not printed for a stated output ilevbare2013c E7
baird2011\Delta G_{fus} inside the activity coefficient of four drugs in PEG 400 at 25 °Cno \Delta G_{fus} value printednot applicable; checked through the activity coefficient (0.614 for chlorpropamide, reproduces) [baird2011 E6, E7]
raina2015aΔG in the amorphous solubility estimate of six dihydropyridinesΔHf and Tm not printedinputs not printed raina2015a E2
ilevbare2013aΔG inside Eq. 1 for eight drugsritonavir 10.1 kJ/mol at 37 °C from Table 2 inputsno ΔG printed; recomputed ilevbare2013a E2
almeidaesousa2015comparator of the Cp method3.24–10.33 kJ/mol for six drugsreproduces (ibuprofen 3.25 against 3.24 kJ/mol) almeidaesousa2015 E2, R11
indulkar2015ΔG at 310 K for clotrimazole and atazanavir6.26 kJ/mol (clotrimazole, with an unprinted molar mass)inputs not printed indulkar2015 E3
mosqueragiraldo2015telaprevir13.31 kJ/mol; amorphous solubility 245 ± 5 µg/mLreproduces mosqueragiraldo2015 E1, E2
almeidaesousa2016CRH15.68 kJ/molreproduces (5.685) [almeidaesousa2016 E2]
indulkar2017aΔG of atazanavir inside Eq. 314.55 kJ/mol from Table I (ΔHf 90.1 J/g, Tm 208 °C, 310 K)not printed; with 0.22 read as exp[−I(a)] it gives the printed 62, and about 228 read as I(a) (synthesis) [indulkar2017a E2, X1; indulkar2015 M2]
hsieh2012amorphous solubility of five basesΔHf, Tm not printedinputs not printed hsieh2012 E3
ilevbare2012britonavirΔHf 60.4 kJ/mol, Tm 121 °C, 37 °CΔG 10.1 kJ/mol, exp(ΔG/RT) 50.8 (extractor) [ilevbare2012b E9]
hsieh2014glyburide, warfarin175.6 °C and 45.5 kJ/mol; 169.2 °C and 43.6 kJ/molreproduces: 59.9 and 47.7 at 25 °C hsieh2014 E2, E3
jackson2014danazolΔHf 28.9 kJ/mol, Tm 225.4 °CΔG 6.95 kJ/mol; ratio 15.4 against 19.6 printed [jackson2014 E1, E2]
duong2022bposaconazole, 310 K\Delta G = \Delta H_f(T_m - T)T/T_m^{2}Tm 442.25 K, ΔHf 76.95 J/g; ΔG 16.13 J/g by the extractor duong2022b E3, X2
rumondor2009cdriving force of six drugs tested as a predictor of moisture-induced separation4.1 (ketoprofen) to 11.0 kJ/mol (pimozide); nifedipine 8.2, felodipine 6.1reproduces (nifedipine 8.23, pimozide 10.97, ketoprofen 4.14) [rumondor2009c E3, M1, M2, M5]
rumondor2009bnamed as the pure-drug driving forcenonenamed only [rumondor2009b E1]
konno2008afelodipine amorphous solubility, 37 °Cratio about 10, about 9.5 µg/mLinputs not printed [konno2008a E2, R17]
alonzo2010felodipine, 37 °C30.12 kJ/mol, 143.41 °C: ΔG 5729 J/mol, ratio 9.22 against about 9reproduces alonzo2010 E1
alonzo2010indomethacin, 37 °C; 25 °C32.48 kJ/mol, 153.4 °C: ΔG 6445 J/mol, ratio 12.17, 36.5 against about 37 µg/mL; 15.8–23.6 against printed 17–25 µg/mLreproduces; mismatch at 25 °C alonzo2010 E2, X2
alonzo2011felodipine, 37 and 25 °Cabout 9.1 and 5–6 µg/mLinputs not printed [alonzo2011 E3, M1]
duong2022adelamanid amorphous-to-crystalline solubility ratio at 37 °C, against the measured 42.3ratio 31.8; Tm 196.1 °Cinputs partly printed; (synthesis: a ratio of 31.8 at 310 K needs ΔG ≈ 8.9 kJ/mol, so ΔHf ≈ 39.8 kJ/mol) duong2022a E2, R27, M1

Where it fails or is disputed

The ΔGv term does not rank the classes monotonically: class II, not class III, has the most positive contribution vaneerdenbrugh2010a X10. A printed ΔGv in one table does not follow from its own inputs baird2012 X2. Similar viscosity with a twofold different ΔGv weakens a viscosity-only comparison baird2012 X9. For vemurafenib the larger driving force of α did not make α dominate at 120–190 °C lu2016 C52. Outside this chapter, errors at large undercooling are reported almeidaesousa2015 E2; mosqueragiraldo2015 E2. In the PEG dispersions paper the relation is evaluated at 25 °C, far below the melting points of 126.7–151.7 °C, and the result is the activity coefficient of F3 of the PEG dispersions chapter [baird2011 E7, M1, M2, M3, M4]. In the solution-methods papers, for diverse drugs the heat-capacity method gave estimates much closer to measured LLPS onsets almeidaesousa2015 C43; the Hoffman-based danazol estimate (16 µg/mL) was above the measured 13 µg/mL [jackson2014 R5]. In the amorphous solubility papers it lacked accuracy for diclofenac (28.9 against 52.2 measured ratio) and telaprevir (2.7 times the onset) almeidaesousa2015 R13, R5; mosqueragiraldo2015 R17; Table 10 of the six-drug paper does not predict the size of the error almeidaesousa2015 X8. The DSC method and some inputs are not stated hsieh2014 X11; hsieh2012 X14. The danazol ratio of 19.6 cannot be reproduced [jackson2014 X1]. The temperature of the warfarin and glyburide calculation is not printed (25 °C assumed) hsieh2014 E2. In the physical-stability papers the driving force did not sort the six drugs by moisture-induced separation [rumondor2009c C32]; for nifedipine against felodipine it pointed the right way for nucleation (F2 of the physical stability chapter) marsac2006a R4.l. The 25 °C indomethacin range of the 2010 dissolution paper does not reproduce from its inputs alonzo2010 X2. Felodipine concentrations of 14.1 µg/mL measured by UV exceed the 9.5 µg/mL estimate of the same 2008 paper [konno2008a X5]. For delamanid the estimate lies about a quarter below the measured ratio (42.3), which the authors call "reasonable agreement" duong2022a R27, C43.

Concepts

Hoffman equation, thermodynamic driving force for crystallization, enthalpy of fusion, degree of supercooling, activity coefficient, amorphous-crystalline free energy difference, amorphous solubility advantage. Findings. Crystallization tendency: F6, F7, F13. PEG dispersions: F3. Solution methods: F5. C. Amorphous solubility and LLPS: F5, F7. Crystallization from solution: F2, F16. Physical stability: F2, F13. Supersaturation from ASD dissolution: F5, F7, F23.

← Concepts and formulas

formula · as used in Crystallization tendency

induction time decomposition

Expression

t_{ind} = t_n + t_g ilevbare2013c E2

t_{ind} \propto J^{-1} ilevbare2013c E3

Both relations are printed in the same form in the celecoxib dispersion papers [xie2015 E1, E2; xie2016a E4].

Variables and units

symbolmeaningunit
t_indmeasured induction time, from creating the supersaturation until crystals are detectedmin
t_ntime to form stable nucleimin
t_gtime for nuclei to grow to detectable sizemin
Jnucleation ratem⁻³ s⁻¹

Assumptions and valid domain

The proportionality holds if steady-state nucleation is reached quickly and t_n \gg t_g; nuclei are detectable only after growth ilevbare2013c C14, E2. The relation was not tested for the three drugs or the polymers ilevbare2013c X14. In the celecoxib papers the proportionality is stated as an assumption ("If it is assumed") xie2015 C41; the onset is read from the abrupt rise of extinction at 350 nm together with the fall of absorbance at 249 nm [xie2016a C9].

Use in the library

paperused forvalues or fitted parametersstatus / recomputed
ilevbare2013creading polymer effects on induction time as nucleation-rate effectsinduction times about 2.4–855.5 mininputs not printed ilevbare2013c E2, E3
xie2015celecoxib dispersions (ASD dissolution chapter); celecoxib 22 µg/mL with 5 µg/mL polymers, pH 6.8, 37 °Cdecomposition printed as E1; proportionality as E2; induction within 5 min without polymer, ≈1 h PVP-K12, ≈2 h PVP-K29/32, more than 8 h HPMCAS, HPMCprinted xie2015 E1, E2; no t_n or t_g printed xie2015 E1, R1
xie2016acelecoxib with PAA and cellulose derivatives; with HPMCASdecomposition printed; 960 min taken as the induction time with HPMCAS, an underestimateprinted [xie2016a E4]; no component times printed [xie2016a E4, C39]
chen2015t_ind = t_n + t_g, not separated11 drugsprinted chen2015 E1, C5
li2016asame, with t_n >> t_g assumedthree drugs, 13 bile saltsprinted in words [li2016a E4]
mosqueragiraldo2016T_ind = T_n + T_growthtelaprevir, 26 polymersprinted [mosqueragiraldo2016 E1]
lu2017t_ind = t_n + t_g, read as nucleation when t_n dominatestelaprevir, six bile saltsprinted lu2017 E3, C9
trasi2015t_j = t − t_g with t_g fittedacetaminophen: t_g 7.8–928 sfitted trasi2015 E4, R3
indulkar2018lecithin, bile salt and SIF media, four drugsinduction time as first scattering rise; components not separatednot applicable [indulkar2018 E5]
lu2018telaprevir in bile salt mixtures and complete mediat_{ind} = t_n + t_gnot applicable [lu2018 E2]
indulkar2019batazanavir in SDSadditive may act on either partnot applicable [indulkar2019b E5, C29]
enright2020atazanavir in phospholipid-digest and volunteer mediaprinted in wordsnot applicable [enright2020 E1]
elkhabaz2021atazanavir, posaconazole in six mediaT_{ind} = T_{crit} + T_{grow}, read as nucleation onsetnot applicable [elkhabaz2021 E1]
que2018unseeded paclitaxel at S ≈88 with and without 20 µg/mL HPMCASwithin 6 min against more than 16 hno component times printed; growth on seeds continued in the same system [que2018 R3, C30]

Where it fails or is disputed

A polymer that slows early growth also lengthens t_{ind}, and for strong inhibitors the extinction rises above the baseline before the fitted onset (about 25 against 31.5 min; about 180 against 260 min), so values depend on where the lines are drawn ilevbare2013c X14, R6. The detection method changes the time (SHG earlier than a probe ratio) [jackson2014 R25]. The seeded and unseeded comparison shows that for STC and SGC the nucleation part dominates but for STCDC and SGCDC growth does [lu2017 R10; lu2017a R14, C37]; in the acetaminophen study the induction time is the time to massive nucleation trasi2015 C32. In the SDS study crystal size and agglomeration change across the same threshold as the induction time, so a change in the growth term cannot be excluded [indulkar2019b X7, X5]. Neither celecoxib paper separates t_n from t_g [xie2015 E1; xie2016a E4] (synthesis: growth was slowed about 70-fold with HPMCAS [xie2016a R12], so part of the longer induction time may come from t_g). For paclitaxel the polymer that lengthened the unseeded induction time beyond 16 h only slowed growth on seeds, so the induction time of an unseeded test says little about a dispersion that carries crystals [que2018 R3, R14].

Concepts

nucleation induction time, nucleation rate, nucleation inhibition. Findings. Crystallization from solution: F3, F5, F13, F14, F27, F28, F29, F31, F32, F33, F35, F38, F39. Supersaturation from ASD dissolution: F4, F6, F20, F22.

← Concepts and formulas

formula · as used in Crystallization tendency

initial growth rate from the desupersaturation slope

Expression

R_g = \text{slope of } C \text{ against } t \text{ over the first 2 min} [ilevbare2012a E4]

C_t = -R_g\,t + b ilevbare2013b E8

Variables and units

symbolmeaningunit
R_ginitial bulk crystal growth rateµg mL⁻¹ min⁻¹ or µg min⁻¹
C, C_tdissolved drug concentration or amount (in 50 mL) at time tµg/mL or µg
ttime after seedingmin
bintercept (starting amount)µg

Assumptions and valid domain

Desupersaturation reflects growth of the seeds, and the curve is linear over the first 2 min [ilevbare2012a C14; ilevbare2013b E8]. For felodipine the slope is taken over 5 min [alonzo2012 P4]. Tested for ritonavir, celecoxib and efavirenz at 37 °C and felodipine at 25 °C.

Use in the library

paperused forvalues or fitted parametersstatus / recomputed
ilevbare2012aritonavir growth rates and ratios1.56 µg mL⁻¹ min⁻¹ at 10 µg/mL (text), about 1.4 (figure)concentration–time curves not printed [ilevbare2012a E4, X5]
ilevbare2013bcelecoxib at 10 µg/mL, no polymer and with CA 320S Sub 0.63−69.80 t + 487.46; −11.88 t + 482.40 µgreproduces (≈488 to ≈348 µg in 2 min gives ≈70 µg min⁻¹) ilevbare2013b E8, R8
schram2015slope of concentration against time over 1 h taken as the growth ratefelodipine S 8[schram2015 C5, E4]
lu2017ainitial slope over 30 min (δ 2.4) or 10 min (higher δ), in µg/min for 50 mL5.4, 58.6, 89.2 µg/min in bufferplot readings; per vessel [lu2017a R2, X12]

Where it fails or is disputed

The same no-polymer condition gives 71.0, 69.80 and about 68 µg min⁻¹ in three places of the celecoxib paper, outside the printed standard deviations ilevbare2013b X7. At near-amorphous-solubility concentrations the slope may include secondary nucleation or a drug-rich phase [ilevbare2012b X18]. Seed mass, volume and methanol fraction are often not printed [ilevbare2012c X16].

Concepts

desupersaturation, crystal growth rate. Findings. F5, F7, F23, F33.

← Concepts and formulas

formula · as used in Crystallization tendency

Jackson interface-kinetics growth rate

Expression

U = \frac{2 k_B T}{\pi \lambda^2 \eta}\, f \left[1 - \exp\left(-\frac{\Delta H\,\Delta T}{k_B T\,T_m}\right)\right] \exp\left(-\frac{\Delta S}{k_B}\right) trasi2014a E1.

Variables and units

symbolmeaningunits
Ucrystal growth ratenot stated trasi2014a E1
\lambdaaverage diffusion jump distancenot stated
\etaviscosityPa·s (cited data) trasi2014a P7
ffraction of interface sites that are active growth sitesdimensionless
\Delta H, \Delta Senthalpy and entropy difference between liquid and crystalnot stated
T, T_m, \Delta Ttemperature, melting temperature, undercoolingK

Assumptions and valid domain

Stokes–Einstein holds and viscosity describes diffusion at the interface in the studied range trasi2014a C20; not near Tg, where diffusion decouples from viscosity trasi2014a C21. The entropy term reflects the fraction of molecules with the right orientation to join the crystal trasi2014a C31.

Use in the library

paperused forvalues or fitted parametersstatus / recomputed
trasi2014aframework for the driving-force correction and the entropy fitnone fitted directlyinputs not printed (λ, f and η(T) not given) trasi2014a E1, R10, R11

Where it fails or is disputed

At similar viscosity the drugs fell into two sets whose corrected growth rates differ by about 2–3 orders of magnitude, so factors beyond viscosity and driving force contribute trasi2014a C30. The 2012 viscosity study used a related screw-dislocation form without the entropy term baird2012 E3.

Concepts

crystal growth rate, melt viscosity, entropy of fusion, thermodynamic driving force for crystallization. Findings. F2, F6.

← Concepts and formulas

formula · as used in Crystallization tendency

Kubota-Mullin model

Expression

\frac{R}{R_0} = 1 - \alpha\,\theta_{eq} [alonzo2012 E5; ilevbare2012b E7]

A model of growth inhibition by an adsorbed impurity based on step pinning: the growth rate with impurity relative to that without falls linearly with the surface coverage, where α is an effectiveness factor that depends on the surface energy, the molecular area, the supersaturation and the spacing of adsorption sites, \alpha = \gamma a/(kT\,(\ln S)\,L) [alonzo2012 E5, E7; ilevbare2012b E7]. It was developed for inorganic systems; Alonzo and co-workers state that, to their knowledge, it had not been used for polymers on organic pharmaceutical crystals [alonzo2012 C10]. The authors extend its proportionality between step, face and mass growth-rate ratios to bulk de-supersaturation rates [alonzo2012 C4, E4]. Both felodipine–HPMCAS papers print V_P/V_0 = 1 - \rho_c/l and apply it to AFM spacings: with l 68.5 nm and ρc 3.98 nm it gives 0.94, and with l 15.8 nm it gives 0.75, against measured rate ratios of 2.7 and about 12 [schram2015 E3; schram2016 E4, R12, R14, X3]. (This page also holds the former concept page "Kubota-Mullin model", whose content is about the same equation.)

Variables and units

symbolmeaningunit
R, R_0overall mass growth rate with and without polymerµg mL⁻¹ min⁻¹
αimpurity effectiveness factordimensionless
θ_eqequilibrium surface coverage by adsorbed polymerdimensionless

Assumptions and valid domain

Langmuir-type adsorption (isotherm fit, here shifted by hand for felodipine); the proportionality between step, face and overall mass growth-rate ratios extends to bulk de-supersaturation rates; constant seed surface area and face-area ratios; applies only below the kinetic roughening transition [alonzo2012 C4, C5, C6, C23; ilevbare2012b E7]. How the parameters are obtained: seeded growth rates with and without polymer at several polymer concentrations give R/R_0; the coverage \theta_{eq} comes from an adsorption isotherm fit (modified Langmuir, Langmuir–Freundlich); α is chosen so that the line fits the measured ratios at one supersaturation [alonzo2012 P9; ilevbare2012b P11, E6].

Use in the library

paperused forvalues or fitted parametersstatus / recomputed
alonzo2012felodipine with HPMC at S 8α 0.4 (text, table) or 0.45 (Fig. 5 caption)reproduces with 0.45: 1 − 0.45 × 0.631 = 0.716 against 0.71 plotted; mismatch with 0.4 [alonzo2012 E5, X1]
alonzo2012felodipine with HPMC, α at S 3, 5, 8, 10, 120.7, 0.5, 0.4, 0.6, 0.2 (25 °C)as printed [alonzo2012 R9]
alonzo2012plateaus of R/R00.65 (S 5) and 0.55 (S 10) quotedmismatch: 1 − α at θ = 1 from Table 1 gives 0.5 and 0.4 [alonzo2012 E5, X2]
ilevbare2012britonavir with CAP Adp 0.85, σ 0.99, 37 °Cα = (1 − 0.80)/0.16 = 1.25, θ 0.16, R/R0 0.80 (pH 3.8); α = 1/0.23 = 4.35, θ 0.23, R/R0 0 (pH 6.8)reproduces [ilevbare2012b E7, R16]
ilevbare2012b, alonzo2012reading of αα about 1: adsorbed molecules one critical-nucleus diameter apart at full coverage; above 1: closer, so growth can stop below full coverageinterpretation [ilevbare2012b E13; alonzo2012 C25]
schram2015R_P/R_0 = 1 − ρc/l with l the globule spacingpH 3 l 44 nmqualitative [schram2015 E3, C18]
schram2016same with measured spacing and critical radius, HPMCAS on buffer-grown and dispersion-grown felodipine seeds (S 4, 25 °C); see the pinning formulal 68.5 and 15.8 nm; ρc 3.98 nm; predicted rate ratios about 1.06 and 1.34recomputed [schram2016 E4, R12, R14, X3]

Where it fails or is disputed

α at pH 6.8 was derived from the point it is compared with and is a lower bound; the low-concentration points disagree (0.44–0.09 measured against 1.00–0.96 calculated) [ilevbare2012b X8, R15]. The coverage of the felodipine fit is model-dependent [alonzo2012 X12]. The model was not tested above σ_c [ilevbare2012b X11]. With the printed l and ρc the model gives much smaller effects than measured [schram2016 X3]. The felodipine α at S 8 is 0.4 in text and table and 0.45 in the Fig. 5 caption; the plateaus quoted "from the alpha estimates" do not follow from Table 1; the coverage was shifted by hand [alonzo2012 X1, X2, X12]. The ln S in eq 7 replaces the low-supersaturation approximation S − 1 of the original [alonzo2012 C7]. The arrest criterion is stated as l ≤ ρc and as l ≤ 2ρc, and ρc was not estimated in the 2015 paper [schram2016 X1; schram2015 X7].

Concepts

impurity effectiveness factor, surface coverage; based on step pinning; related to polymer adsorption. Related formulas. Kubota-Mullin impurity effectiveness factor, Langmuir-Freundlich isotherm, proportionality of step velocity and growth rate ratios. Findings. Crystallization from solution: F9, F10, F11, F23, F24.

← Concepts and formulas

formula · as used in Crystallization tendency

linear crystal growth rate from size-time slope

Expression

Printed in words: the growth rate is the slope of the straight line fitted to the linear region of a crystal size–time plot, and the same rule gives an overall crystallization rate from the linear part of a % crystallinity–time plot kestur2013b E2. Printed in words in a growth-rate survey of the library: the slope of crystal length against time trasi2014b E1. In symbols (synthesis):

G = \frac{\Delta L}{\Delta t}

Printed in words in two later papers: growth rate as the slope of spherulite size against time [sato2015 E1]; slope of crystal diameter against time sato2016 E3.

Variables and units

symbolmeaningunits
Glinear (radial or diameter) growth ratem/s or µm/s kestur2013b E2; trasi2014b E1
Lcrystal size (radius, diameter or length)m or µm trasi2014b E1
ttimes trasi2014b E1
crystallization slope% crystallinity per unit time, printed as day⁻¹ or h⁻¹kestur2013b E2, X4

Assumptions and valid domain

Size grows linearly with time; for powders, the early and late stages are each linear kestur2013b E2. At low temperature the radius change can be taken from the percentage of area crystallized instead trasi2014b E1.

Use in the library

paperused forvalues or fitted parametersstatus / recomputed
kestur2013bform I growth at the free surface and in the bulk, 30 °C2.7 × 10⁻⁹ and 0.47 × 10⁻⁹ m/s; 1.45 × 10⁻⁹ and 6.37 × 10⁻¹¹ m/s with 3% PVPsingle values, no uncertainty kestur2013b R12, X11
kestur2013bearly and late crystallization slopes of powders0.905 and 0.072 day⁻¹ (size 1); 0.43 and 0.03 day⁻¹ (size 2)reproduces 0.90 day⁻¹ from plot readings; fitted range not printed kestur2013b R14, E2
kestur2013bearly slope, pure drug against 3% PVP2.21 against 0.037 h⁻¹; 1.41 against 0.018 h⁻¹PVP values equal Table 2 divided by 24 kestur2013b R15, X4
trasi2014b (outside)radial growth of nilutamide and other drugsrates per temperatureinputs not printed trasi2014b E1
sato2015growth of ATP enantiomers with five polymerssize–time lines through the origin, e.g. at 60 °C with HPMCAS-HFlinear in time [sato2015 R4, R7, R8]
sato2016growth of six compounds with PHBdiameter rates; griseofulvin with 5% PHB about tenfold faster at 120 °Cdiameter rate = 2 × radial rate sato2016 R5, R6, X8

Where it fails or is disputed

The flutamide growth paper reports diameter rates, the acetaminophen paper radial rates, a factor of two apart trasi2012b R3; trasi2012a R6. A ratio of powder slopes (% per day) is not the same quantity as a ratio of film growth rates (m/s), although the two were compared kestur2013b X6. The PHB paper takes diameter slopes, twice the radial front speed sato2016 X8.

Concepts

crystal growth rate, nucleation and crystal growth, surface-enhanced crystal growth. Findings. F3, F17, F18, F21, F27, F28.

← Concepts and formulas

formula · as used in Crystallization tendency

linear fit of melt viscosity on Tm − Tg

Expression

y = -0.0028\,x + 0.3989,\quad R^2 = 0.7368 trasi2014a E6.

Variables and units

symbolmeaningunits
yviscosity at TmeltPa·s trasi2014a E6
xT_m - T_gK trasi2014a E6

Assumptions and valid domain

None stated trasi2014a E6. Fitted on the 17 compounds with a melt viscosity cited from the 2012 viscosity study, Tm − Tg about 85–146 trasi2014a R9.

Use in the library

paperused forvalues or fitted parametersstatus / recomputed
trasi2014ashowing that slow growers have high melt viscosity and small Tm − Tgslow group at 85–117 with 0.0703–0.191 Pa·s; fast group at 121–146 with 0.0136–0.0513 Pa·sreproduces: slope −0.00277, intercept 0.3991, R² 0.7369 trasi2014a E6, R9

Where it fails or is disputed

The authors describe a trend "with considerable scatter" trasi2014a C26; three group 2 compounds fall among group 1, explained by fragility and complexity that were not measured trasi2014a C29, X18. One Tm − Tg value in the source table does not match its own Tm and Tg trasi2014a X1. The fit reaches zero at x = 0.3989/0.0028 = 142.5 K and gives −0.0099 Pa·s at x = 146 (0.3989 - 0.0028 \times 146), so it is not usable at the upper end of its own data (synthesis, from trasi2014a E6, R9).

Concepts

melt viscosity, Tm − Tg temperature gap, crystal growth rate. Findings. F5.

← Concepts and formulas

formula · as used in Crystallization tendency

linear fit of normalized growth rate on melting entropy

Expression

y = -0.6\,x - 0.95,\quad R^2 = 0.61 trasi2014a E7.

Variables and units

symbolmeaningunits
y\log(U_{kin}\,\tau_\alpha/a), growth rate corrected for driving force and normalized by relaxation time and molecular sizedimensionless trasi2014a E7, E2, E4
x\Delta S_m/Rdimensionless trasi2014a E7

Assumptions and valid domain

None stated trasi2014a E7; τ_α is taken from viscosity as \tau_\alpha = \frac{C\eta}{G} trasi2014a E4, which assumes Stokes–Einstein behaviour away from Tg trasi2014a C20, C21. Nine compounds with viscosity–temperature data trasi2014a R11.

Use in the library

paperused forvalues or fitted parametersstatus / recomputed
trasi2014atesting whether melting entropy explains growth differences at equal mobilityslope −0.6, intercept −0.95, R² 0.61reproduces from the nine plotted points: slope ≈ −0.61, intercept ≈ −0.91, R^2 \approx 0.61 trasi2014a E7, R11

Where it fails or is disputed

The decrease follows expectation but with "considerable scatter"; entropy "certainly plays a role" while other factors, possibly interactions in the melt and at the interface, are not accounted for trasi2014a C32.

Concepts

entropy of fusion, crystal growth rate, melt viscosity. Findings. F6.

← Concepts and formulas

formula · as used in Crystallization tendency

power-law crystal growth rate

Expression

R_G = k_G\,A\,(C_b - C^{*})^{g} [purohit2018 E3]. The log-linearised fit \log(dC/dt) = n\log(S-1) + \text{intercept} is printed for tacrolimus seeds, with n = 1.677 [hate2019 E5].

In supersaturation-ratio form, with its linearisation, and against the logarithmic supersaturation:

R_G = k_G\,S^{g} [alonzo2012 E2];

\ln R_G = \ln k_G + g\,\ln S [alonzo2012 E3];

R_G = k_g\,\sigma^{g} [ilevbare2012b E3].

Variables and units

symbolmeaningunits
R_Gcrystal growth rate–
k_Ggrowth rate coefficientnot determined
Acrystal surface areanot determined
C_b, C^*bulk concentration, crystal solubilityµg/mL
goverall growth order, typically 1 to 2–
R_G (supersaturation form)overall crystal growth rateµg mL⁻¹ min⁻¹ [alonzo2012 E2]
k_ggrowth rate constant against σas R_G [ilevbare2012b E3]
S, \sigmasupersaturation ratio and its logarithmdimensionless

Assumptions and valid domain

Empirical [purohit2018 E3]; g = 1 for diffusion control and 2 for integration control in the felodipine–HPMC study [alonzo2012 E2, E3]. k_G depends on temperature, crystal–solution velocity and impurities but not on supersaturation [alonzo2012 C3, C35]. Fitted over S 3–12 for felodipine; not fitted for ritonavir [alonzo2012 P8; ilevbare2012b E3].

Use in the library

paperused forvalues or fitted parametersstatus / recomputed
purohit2018why 450 mL shows growth and 900 mL notnone fittedinputs not printed [purohit2018 E3, C52, C55]
alonzo2012growth order of felodipine without and with HPMCg 1.50 ± 0.04; 1.97 ± 0.12; \ln k_G −4.24; −5.42 to −5.86; with HPMC kG about 0.0032, g 2, S 5: RG about 0.080reproduces (1 µg/mL HPMC, S 5: 0.080 against 0.087 µg mL⁻¹ min⁻¹ in Fig. 3) [alonzo2012 E2, E3]
hate2019tacrolimus on seedsdC/dt = k_g A (C_{in}(0) - C^*)^n with constant area; fit y = 1.677x - 3.136, R² 0.9896E4 reproduces (1.17 × 10⁻⁴ × (25 − 2.5)^1.7 = 0.0233 µg/mL/s); E5 mismatch: printed growth constant 1.17 × 10⁻⁴ "cm/s", 10^(−3.136) = 7.31 × 10⁻⁴ or 1.57 × 10⁻⁴ after dividing by C*^n [hate2019 E4, E5, R18, X3]
ilevbare2012bgrowth against σ; linear above σ_cno k_g or g reportednot applicable [ilevbare2012b E3]
schram2015R_G = k_G S^g, held at fixed disc speedS 8; no k_G or g reportednot fitted [schram2015 E2, X7]
schram2016R_G = A k_G S^g, area factor in the argument against area as the causeS 4; no k_G or g reportednot fitted [schram2016 E2]

Where it fails or is disputed

No growth constants were determined for tacrolimus in the dissolution study [purohit2018 E3]; the absorptive study fitted an order and a constant from seeded runs. The x-axis of the tacrolimus fit is log(C − C*) rather than the log(S − 1) it is labelled with, and the crossover with absorption near S 18.5 rests on a constant crystal area (F29) [hate2019 X2, X16]. The k_G values come from fits with different fixed orders (1.5 without HPMC, 2 with), so they should be compared at equal S [alonzo2012 X8]. Above the critical supersaturation the ritonavir rate is linear in σ, not a power law with order 2 [ilevbare2012b R11, R12]. The rotating-disc papers of 2015 and 2016 report neither order nor coefficient [schram2015 X7; schram2016 E2].

Concepts

crystal growth, crystal seeding, crystal growth rate, integration-controlled crystal growth, kinetic roughening. Findings. Bioperformance: F12, F16, F29. Crystallization from solution: F9, F10, F23, F24.

← Concepts and formulas

formula · as used in Crystallization tendency

reduced glass transition temperature

Expression

T_{rg} = \frac{T_g}{T_{melt}} baird2010 E4.

Variables and units

symbolmeaningunits
T_{rg}reduced glass transition temperaturedimensionless baird2010 E4
T_g, T_{melt}glass transition and melting temperaturesK baird2010 E4

Assumptions and valid domain

Viscosity is taken as constant at Tg baird2010 E4; a value of 0.66 was proposed in the cited literature baird2010 R29. For 16 class I molecules without a measured Tg, Tg was estimated from a fit T_{rg} = 0.00021\,\mathrm{MW} + 0.64433 with R² 0.44 baird2010 E27, X17.

Use in the library

paperused forvalues or fitted parametersstatus / recomputed
baird2010testing Trg against classI-A 0.69, I-B 0.7; class I 0.69 ± 0.02, class III 0.73 ± 0.04, not significantly differentreproduces: acetaminophen 297.15/443.15 = 0.671 baird2010 E4, R14
nguyen2023apretomanid0.67reproduces: (10 + 273.15)/(147.3 + 273.15) = 0.673 nguyen2023a E2

Where it fails or is disputed

Trg did not differ between class I-A and I-B, indicating no correlation with the ability to quench amorphous baird2010 C23. Tg/Tm was not a reliable indicator; class II alone spans Tg/Tm 0.65–0.76 baird2012 C36, R14. The class I values rest partly on estimated Tg baird2010 X17.

Concepts

reduced glass transition temperature, glass transition temperature, glass-forming ability, Tm − Tg temperature gap. Findings. F12.

← Concepts and formulas

formula · as used in Crystallization tendency

relative nucleation rate from seeded and unseeded induction times

Expression

\frac{R}{R_0} = \frac{t_{s,0}}{t_s} [alonzo2012 E9]

\frac{J}{J_0} = \left(\frac{t_s}{t_{s,0}}\right)^{3}\left(\frac{t_{u,0}}{t_u}\right)^{4} [alonzo2012 E10]

\frac{J}{J_0} = \left(\frac{R_0}{R}\right)^{3}\left(\frac{t_{u,0}}{t_u}\right)^{4}, \qquad E_J = \frac{J_0}{J} [lu2017 E4; lu2017a E4, E5]

With the polymer case in the denominator:

\frac{J_0}{J_p} = \left(\frac{R_p}{R_0}\right)^{3}\left(\frac{t_{u,p}}{t_{u,0}}\right)^{4} [xie2016a E6]

Variables and units

symbolmeaningunit
J, J_0nucleation rate with and without inhibitorm⁻³ s⁻¹
t_s, t_s,0seeded induction times with and without inhibitormin
t_u, t_u,0unseeded induction times with and without inhibitormin
R/R_0seeded growth-rate ratio, used for t_{s,0}/t_sdimensionless
J_0, J_pnucleation rate without and with polymerm⁻³ s⁻¹
R_0, R_pmass growth rate without and with polymer, from seeded desupersaturation slopesµg/(mL min)
t_{u,0}, t_{u,p}unseeded induction times without and with polymermin

Assumptions and valid domain

Seeded and unseeded runs under identical conditions; the seeded induction-time ratio is replaced by the growth-rate ratio of the de-supersaturation curves (Verdoes 1992, outside the library) [alonzo2012 C11, E9]. Used for one drug, one polymer level and one supersaturation. For celecoxib R_0 is the slope of the linear region over the first 5 min without polymer; R_p is, for simplicity, the slope over the whole 16 h with HPMCAS [xie2016a C38]. t_{u,p} = 960 min because no sharp slope change was seen, which underestimates the induction time [xie2016a C39].

Use in the library

paperused forvalues or fitted parametersstatus / recomputed
alonzo2012felodipine, 1 µg/mL HPMC, S 10t_{u,0} 15.3 min, t_u 136.5 min, R/R_0 0.607; J/J_0 = 7.05 × 10⁻⁴reproduces: 7.06 × 10⁻⁴ with 1.647; 7.00 × 10⁻⁴ with the printed 1.643 [alonzo2012 E10, X6]
xie2016acelecoxib with HPMCAS (ASD dissolution chapter): 22 µg/mL with 5 µg/mL HPMCAS, pH 6.8, 37 °Ct_{u,p} 960 min, R_0/R_p = 70; J_0/J_p about 4,000derived [xie2016a R13]; reproduces with t_{u,0} = 5 min from the text: (1/70)^{3}(960/5)^{4} = 3962; the t_{u,0} used by the authors is not printed [xie2016a E6, R13]
lu2017telaprevir, 1.86 mM STC and SGC, δ 11.5; t_u,0 about 30 min, t_u 960 minJ/J_0 2.5 × 10⁻³ and 2.9 × 10⁻⁶recomputed 2.58 × 10⁻³ and 3.06 × 10⁻⁶ lu2017 E4, R10, X1
lu2017asame with bile salts at 1.86 and 12 mM; rate ratio set to 100 for STCDC and SGCDCE_J about 360 to 3 × 10⁵; about 1 for STCDCreproduced for the 1.86 mM values [lu2017a E4, E5, R14]

Where it fails or is disputed

Rests on one pair of induction times with no stated uncertainty entering to the fourth power; a 10% change in t_u changes J/J_0 about 1.5-fold [alonzo2012 X7]. The exponents are not derived in the paper. The induction times enter capped at 960 min, so E_J is a bound; the rate ratios used are the δ 12.5 values while the text speaks of growth; the 12 mM inputs are not printed [lu2017 X2; lu2017a X13, X14]. For celecoxib the slopes and t_{u,0} are not printed, and R_p was taken over 16 h while the figure covers 4 h, so the ratio cannot be reproduced from the printed data [xie2016a X8]. With t_{u,p} capped at 960 min the ratio is a lower bound [xie2016a C39, C40]. t_{u,0} enters to the fourth power (synthesis: with t_{u,0} of 4 or 6 min the ratio is about 9,700 or 1,900).

Concepts

nucleation rate, nucleation inhibition, crystal growth inhibition. Findings. Crystallization from solution: F5, F6, F20, F32, F33. Supersaturation from ASD dissolution: F5.

← Concepts and formulas

formula · as used in Crystallization tendency

solid-liquid equilibrium solubility equation

Expression

\ln x_{C,L_1} = -\frac{\Delta G_{a\to c}}{RT} - \ln\gamma_{L_1} ilevbare2013a E3;

\ln C_B = -\frac{\Delta G}{RT} - \ln\gamma indulkar2015 E7;

and for the supercooled liquid \ln x_{SL,L_1} = -\ln\gamma_{L_1} ilevbare2013a E4. As printed, \Delta G_{a\to c} and \Delta G are positive magnitudes; with the signed quantity \mu^{(c)}-\mu^{0(a)}, which is negative, the term carries a plus sign, as in Taylor and Zhang 2016 (Eqs. 2 and 6).

New forms (Miscibility and drug-polymer interactions). With the heat-capacity terms, as printed for the solubility of a crystalline drug in a polymer (2006):

\ln\gamma_{\mathrm{drug}}\,x_{\mathrm{drug}} = -\frac{\Delta H_{fus}}{RT}\left[1-\frac{T}{T_m}\right] - \frac{1}{RT}\int_{T_m}^{T}\Delta C_P^{\mathrm{config}}\,dT + \frac{1}{R}\int_{T_m}^{T}\frac{\Delta C_P^{\mathrm{config}}}{T}\,dT marsac2006b E5;

and in the 2009 paper, where the left side is written as the free energy of fusion:

\ln\gamma_{\mathrm{drug}}\,x_{\mathrm{drug}} = \frac{-\Delta G_{fus}}{RT} = -\frac{\Delta H_{fus}(T_m)}{RT}\left[1-\frac{T}{T_m}\right] - \frac{1}{RT}\int_{T_m}^{T}\Delta C_P\,dT + \frac{1}{R}\int_{T_m}^{T}\frac{\Delta C_P}{T}\,dT marsac2009 E1.

Simpler forms in other papers of the library:

\ln x_{actual} = -\frac{\Delta H_{fus}}{RT}\left(1-\frac{T}{T_M}\right) - \ln\gamma marsac2008a E3;

\ln\left[\gamma_d\,x_d\right] = -\frac{\Delta G_{fus}}{RT} [baird2011 E5];

\ln x = -\frac{\Delta H_f}{R}\left(\frac{1}{T}-\frac{1}{T_m}\right) - \frac{\Delta C_p}{R}\left(1-\frac{T_m}{T}+\ln\frac{T_m}{T}\right) - \ln\gamma raina2014a E1.

The ritonavir–atazanavir paper writes \log\chi = \log X_{ideal} - \log\gamma with \log X_{ideal} = -\frac{\Delta H_m}{2.303R}\frac{T_m - T}{T_m T} [alhalaweh2016 E4, E5].

Variables and units

symbolmeaningunits
x_{C,L_1}, C_Bsolubility of the crystalline drug in watermole fraction; µg/mL ilevbare2013a E3; indulkar2015 E7
\Delta Gfree-energy difference between supercooled liquid and crystalindulkar2015 E7
\gamma_{L_1}, \gammaactivity coefficient of the drug in the aqueous solutiondimensionless ilevbare2013a E3
\gamma_{drug}activity coefficient of the drug at the solubility limitdimensionless marsac2009 E1
\chi, X_{ideal}mole-fraction solubility and ideal solubility in the ritonavir–atazanavir paper– [alhalaweh2016 E4, E5]
x_{drug}mole-fraction solubilitydimensionless marsac2009 E1
\Delta H_{fus}, T_mheat of fusion and melting temperature of the drugkJ/mol, K (Table I and II values) marsac2006b M1, M2; marsac2009 M1, M2
\Delta C_P^{config}, \Delta C_Pheat-capacity difference between liquid and crystalvalues cited, not printed marsac2006b X11; marsac2009 P7
TtemperatureK; not stated for the Table II predictions marsac2006b X11

Assumptions and valid domain

The solvent is not present in the crystal; the first term is the lattice contribution and the second the solute–solvent interaction indulkar2015 E7; γ is constant over the crystalline and amorphous concentration range ilevbare2013a C55. For a drug in a polymer, the polymer assumes the role of a solvent, so the equation describes solubility of a crystalline drug in a polymer, with \gamma from Flory–Huggins theory marsac2006b C45. For the analogue liquid the entropy of mixing is assumed ideal marsac2006b C46. For ketoconazole the heat-capacity difference was approximated by its value at Tg, integrated between Tg and Tm marsac2009 P7. In the 2009 route the same γ is assumed at the solubility limits in the analogue and in PVP marsac2009 C11. The amorphous form removes the fusion terms, so \ln x = -\ln\gamma raina2014a E2. The ritonavir–atazanavir paper sets ΔCp to zero [alhalaweh2016 E5].

Use in the library

paperused forvalues or fitted parametersstatus / recomputed
ilevbare2013aappendix derivation of the amorphous solubilitynoneγ not printed ilevbare2013a E3, E4
indulkar2015argument that the un-ionized amorphous and crystalline solubility differ by a constant factornoneγ not printed indulkar2015 E7, C42
marsac2006bsolubility of nifedipine and felodipine in PVP K12, PVP K29/32 and methyl pyrrolidone0.06–0.07 g/g and 0.25–0.31 g/g in PVP; 0.15 and 0.48 g/g in methyl pyrrolidonereproduces with ΔC_P terms set to zero at 298.15 K for nifedipine in PVP K12 (0.071 against 0.07); one felodipine value in K29/32 does not match (≈0.29 against 0.25) marsac2006b E5, E6, R17, X11
marsac2009γ in EP from the measured solubility; ideal solubilityγ 0.15–0.89inputs not printed (heat capacities, ideal solubilities) marsac2009 E1, R9, X4
baird2011activity coefficient in PEG 400see the PEG chapterreproduces (through its E7) [baird2011 E5]
marsac2008aactivity coefficient of nifedipine and felodipine in water from the solubility in methyl pyrrolidoneγ in water 1.89 × 10³ and 2.43 × 10⁵reproduces within the stated SD with the lattice terms implied by the printed data (−5.40 and −3.51); ΔH_fus and T_M not printed marsac2008a E3, R10, R11
raina2014abackground to the amorphous solubility predictionnoneinputs not printed (ΔC_p and γ not printed) raina2014a E1, R7
alhalaweh2016crystal and solvation contributions to the solubility of ritonavir and atazanavir at 310 Klog γ about 4.45 and 3.88 (read)mismatch: recomputed 4.65 and 3.01 from the card's ideal solubility and the Table 2 crystalline solubility [alhalaweh2016 E4, R7]
alhalaweh2016ideal-solubility term of ritonavir and atazanavirplotted log X_ideal −1.07, −1.97; recomputed −2.59, −4.53mismatch [alhalaweh2016 E5, X1, X2]
marsac2008anifedipine and felodipine in methyl pyrrolidone and water (second reading)γ in water 1.73 × 10³ and 2.31 × 10⁵reproduces within SD marsac2008a E3

Where it fails or is disputed

The appendix prints its first equation as "(1)" but cites it as Eq. 3 ilevbare2013a X4. The temperature and the heat-capacity values of the 2006 Table II are not printed, the methyl pyrrolidone check is about 30% low for nifedipine, and the polymer-phase values are predictions, not measurements marsac2006b X10, X11; marsac2009 X17. Solubility measured in a small analogue overestimates solubility in the polymer unless corrected for the smaller mixing entropy marsac2006b C51. The plotted terms of the ritonavir–atazanavir paper mix concentration scales; the ordering of the terms survives the recomputation [alhalaweh2016 X2]. For marsac2008a the γ in water is given as 1.89 × 10³ and 2.43 × 10⁵ in one reading and as 1.73 × 10³ and 2.31 × 10⁵ in another; both are reported as reproducing within the stated SD marsac2008a E3, R10, R11.

The ratio form of the same relation for liquid PEG is on the page activity coefficient from solubility.

Concepts

crystalline solubility, activity coefficient, ideal solubility, drug solubility in polymer, micellar solubilization, amorphous solubility. Findings. C. Amorphous solubility and LLPS: F5, F10, F12. Miscibility and drug-polymer interactions: F8, F9. Bioperformance: F1, F6.

← Concepts and formulas

formula · as used in Crystallization tendency

temperature of maximum crystal growth rate

Expression

T_{cryst,max} \approx 0.94\,T_m trasi2014a E8;

T(\text{max growth}) \approx 0.94\,T_m, printed in words as Naito's rule lu2016 E2.

The physical-stability papers give about 0.94 Tm sato2016 E2 and about 0.95 Tm:

T_{max} \approx 0.95\,T_m tian2017 E3.

Variables and units

symbolmeaningunits
T_{cryst,max}temperature of the maximum crystal growth velocityK trasi2014a E8
T_mmelting temperature (of the α-form for vemurafenib)K trasi2014a E8; lu2016 E2
T_{max}temperature of fastest growthK, for the ratio tian2017 E3

Assumptions and valid domain

A cited empirical observation (ref 26) trasi2014a E8, a typical value (ref 21) lu2016 E2. The ratio holds only in kelvin: in °C the vemurafenib product would be 254.7 °C instead of 238.4 °C lu2016 E2. The rule refers to the pure drug. With 10% polymer the temperature of fastest growth of indomethacin moved, for α from 135 to 100 °C (HPMC), 110 °C (HPMCAS) and 125 °C (PVP), and for δ to 105–115 °C, so the rule does not give the maximum of a drug–polymer melt tian2017 R15. In the PHB paper all growth temperatures were placed below 0.94 Tm to read growth as transport-limited sato2016 E2, C15.

Use in the library

paperused forvalues or fitted parametersstatus / recomputed
trasi2014agrouping 31 drugs by maximum growth rateTcryst,max not tabulated; ΔG and ΔSm columns imply 0.90–0.99 Tminputs not printed trasi2014a E8, X4
lu2016expected maximum for α-vemurafenib240 °C predicted against 250 °C observedreproduces: 0.94 × 544.15 K = 511.5 K = 238.4 °C lu2016 E2, R24, C18
sato2016position of measured temperatures below the maximum0.94 Tm above all highest growth temperaturesreproduces sato2016 E2
tian2017indomethacin polymorphs≈0.95 Tm, within the 5 °C step of the datareproduces tian2017 E3
tian2017maxima of indomethacin polymorphs with and without polymerpure drug 110, 135, 135 °C (δ, α, γ); with polymer 100–130 °Cthe rule holds for the pure drug (row above); with polymer the maximum is lower tian2017 R15, E3

Where it fails or is disputed

The 0.94 Tm in the conclusions of the 31-drug study is a cited rule, not a result of that paper trasi2014a X5. The observed vemurafenib maximum of 250 °C corresponds to 523.15/544.15 = 0.96\,T_m (synthesis, from lu2016 R12.o, E2). Inhibition ratios rise steeply above the polymer-lowered maximum (indomethacin α with HPMC about 11 at 100 °C and 2900 at 120 °C), so a ratio taken between the two maxima compares growth in different regimes tian2017 R12, C35.

Concepts

maximum crystal growth velocity, crystal growth rate, degree of supercooling, kinetic versus thermodynamic control of crystal growth. Findings. Crystallization tendency: F1, F2. Physical stability: F3, F28, F29.

← Concepts and formulas

formula · as used in Crystallization tendency

Vogel-Tammann-Fulcher equation

Expression

\tau = \tau_0 \exp\left(\frac{D\,T_0}{T - T_0}\right) baird2010 E12;

\eta = \eta_0 \exp\left(\frac{D\,T_0}{T - T_0}\right) baird2012 E4.

Variables and units

symbolmeaningunits
\tau, \etamean molecular relaxation time, or viscositys; Pa·s baird2010 E12; baird2012 E4
\tau_0, \eta_0value for the unrestricted materialτ0 ≈ 10⁻¹⁴ s baird2010 E12
Dstrength parameterdimensionless baird2012 E4
T_0zero-mobility temperature, for some systems the Kauzmann temperatureK baird2012 E4

Assumptions and valid domain

For the strength parameter D^b: T_0 = T_K, τ(Tg) ≈ 10² s, τ0 ≈ 10⁻¹⁴ s baird2010 E12. Valid from Tm down to about 0.4 Tg on a scale with T_g = 0 and T_m = 1, with D and T0 fitted together baird2012 E4. Strong liquids have D above 30, fragile ones below 10 baird2010 R30.

Use in the library

paperused forvalues or fitted parametersstatus / recomputed
baird2010D^b from Tg and Kauzmann temperature for class II and IIIindomethacin D^b 5reproduces: \ln(10^{16}) \times (318.15 - 282.15)/282.15 = 4.70 baird2010 E12, R18
baird2012fits of melt viscosity for 28 compoundsD 4.3–8.7, all below 10inputs not printed (η0 not reported) baird2012 E4, R12

Where it fails or is disputed

The fitted T0 and D do not satisfy the paper's own relation between T0, D and Tg baird2012 X5. The six class I-A compounds were not fitted, though the abstract calls all compounds fragile baird2012 X7. Whether a compound counts as strong or fragile depends on using m or D baird2010 X14. Good and poor glass formers showed the same non-Arrhenius behaviour, so kinetic fragility "may not accurately assess" glass-forming ability baird2012 C43; fragility was also invoked, untested, for two growth-rate outliers trasi2014a C29, X18.

Concepts

fragility, melt viscosity, glass transition temperature, supercooled liquid. Findings. F5, F12.